

Cp2k: An electronic structure and molecular dynamics software package-quickstep: Efficient and accurate electronic structure calculations
Open accessThe DOI matches its Crossref record; 444 of the 445 references it cites were found in a registry or an index.
Abstract
CP2K is an open source electronic structure and molecular dynamics software package to perform atomistic simulations of solid-state, liquid, molecular, and biological systems. It is especially aimed at massively parallel and linear-scaling electronic structure methods and state-of-the-art ab initio molecular dynamics simulations. Excellent performance for electronic structure calculations is achieved using novel algorithms implemented for modern high-performance computing systems. This review revisits the main capabilities of CP2K to perform efficient and accurate electronic structure simulations. The emphasis is put on density functional theory and multiple post–Hartree–Fock methods using the Gaussian and plane wave approach and its augmented all-electron extension.
Overview
The claims and statistics of this paper haven’t been checked yet.
Free: it reads the open-access text and checks the first 5 claims that cite a source we can check. Create a free account to check every claim.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Free: it reads the open-access text and checks the first 5 claims that cite a source we can check. Create a free account to check every claim.
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Items 401–444 of 444
Golze, D., Iannuzzi, M., & Hutter, J. (2017) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.7b00148 · Cited by 16
Self-citationTumbling with a limp: local asymmetry in water's hydrogen bond network and its consequences
Elgabarty, H., & Kühne, T. D. (2020) · Physical Chemistry Chemical Physics · doi:10.1039/c9cp06960g · Cited by 16 · cited 1×
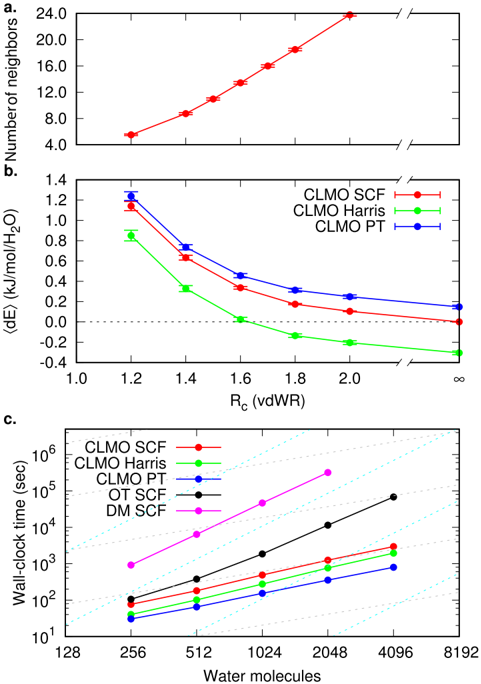
“180,251–256 While the localization constraints are necessary to design orbital-based O ( N ) methods, the reduced number of electronic degrees of freedom results in the variationally optimal CLMO energy being always higher than the reference energy of fully delocalized orbitals.”
BackgroundSelf-citationDensity functional theory study of self-trapped holes in disordered SiO2
Camellone, M. F., Kühne, T. D., & Passerone, D. (2009) · Physical Review B · doi:10.1103/physrevb.80.033203 · Cited by 15
Self-citationYi, Z., Ma, Y., Zheng, Y., Duan, Y., & Li, H. (2018) · Advanced Materials Interfaces · doi:10.1002/admi.201801175 · Cited by 15
On the Possibility of Helium Adsorption in Nitrogen Doped Graphitic Materials
Sahoo, S. K., Heske, J., Azadi, S., Zhang, Z., Tarakina, N. V., Oschatz, M., Khaliullin, R. Z., Antonietti, M., & Kühne, T. D. (2020) · Scientific Reports · doi:10.1038/s41598-020-62638-z · Cited by 15
Self-citationOptimized multi-site local orbitals in the large-scale DFT program CONQUEST
Nakata, A., Bowler, D., & Miyazaki, T. (2015) · UCL Discovery (University College London) · Cited by 15
Valdemoro, C. (1996) · International Journal of Quantum Chemistry · doi:10.1002/(sici)1097-461x(1996)60:1<287::aid-qua29>3.0.co;2-a · Cited by 14
Shi, Y., Scheiber, H., & Khaliullin, R. Z. (2018) · The Journal of Physical Chemistry A · doi:10.1021/acs.jpca.8b06857 · Cited by 14
Self-citationA comparison of accelerators for direct energy minimization in electronic structure calculations
Baarman, K., & VandeVondele, J. (2011) · The Journal of Chemical Physics · doi:10.1063/1.3603445 · Cited by 13
Self-citationGabardi, S., Caravati, S., Los, J. H., Kühne, T. D., & Bernasconi, M. (2016) · The Journal of Chemical Physics · doi:10.1063/1.4950817 · Cited by 13
Self-citationLow-Dimensional Confined Ice Has the Electronic Signature of Liquid Water
Yun, Y., Khaliullin, R. Z., & Jung, Y. (2019) · The Journal of Physical Chemistry Letters · doi:10.1021/acs.jpclett.9b00921 · Cited by 13
Self-citationA hybrid approach to Fermi operator expansionPreprint
Ceriotti, M., Kühne, T. D., & Parrinello, M. (2013) · Cited by 13
Self-citationA Hybrid Approach to Fermi Operator Expansion
Ceriotti, M., Kühne, T. D., Parrinello, M., Maroulis, G., & Simos, T. E. (2009) · AIP conference proceedings · doi:10.1063/1.3225396 · Cited by 12
Self-citationAngerer, C. M., Polig, R., Zegarac, D., Giefers, H., Hagleitner, C., Bekas, C., & Curioni, A. (2015) · Sustainable Computing Informatics and Systems · doi:10.1016/j.suscom.2015.10.001 · Cited by 12
Inverse simulated annealing for the determination of amorphous structures
Los, J. H., & Kühne, T. D. (2013) · Physical Review B · doi:10.1103/physrevb.87.214202 · Cited by 12
Self-citationSchusteritsch, G., Kühne, T. D., Guo, Z., & Kaxiras, E. (2016) · The Philosophical Magazine A Journal of Theoretical Experimental and Applied Physics · doi:10.1080/14786435.2016.1217360 · Cited by 12
Self-citationElgabarty, H., Schmieder, P., & Sebastiani, D. (2012) · Chemical Science · doi:10.1039/c2sc21145a · Cited by 11
Partovi‐Azar, P., & Kühne, T. D. (2015) · physica status solidi (b) · doi:10.1002/pssb.201552236 · Cited by 11
Self-citationLiquid methane at extreme temperature and pressure: Implications for models of Uranus and Neptune
Richters, D., & Kühne, T. D. (2013) · Journal of Experimental and Theoretical Physics Letters · doi:10.1134/s0021364013040127 · Cited by 11
Self-citationPSelInv – A distributed memory parallel algorithm for selected inversion: The non-symmetric case
Jacquelin, M., Lin, L., & Yang, C. (2017) · Parallel Computing · doi:10.1016/j.parco.2017.11.009 · Cited by 11
Proceedings of the Platform for Advanced Scientific Computing Conference
(2016) · doi:10.1145/2929908 · Cited by 11
Declerck, R., Pauwels, E., Speybroeck, V. V., & Waroquier, M. (2008) · The Journal of Physical Chemistry B · doi:10.1021/jp076571w · Cited by 10
Scheiber, H., Shi, Y., & Khaliullin, R. Z. (2018) · The Journal of Chemical Physics · doi:10.1063/1.5029939 · Cited by 10
Self-citationPlessl, C., Platzner, M., & Schreier, P. J. (2015) · Informatik-Spektrum · doi:10.1007/s00287-015-0911-z · Cited by 10
Self-citationSchiffmann, F., & VandeVondele, J. (2015) · The Journal of Chemical Physics · doi:10.1063/1.4922988 · Cited by 9
Self-citationBasis set considerations for the calculation of gradients in the lcao formalism
Almløf, J., & Helgaker, T. U. (1981) · Chemical Physics Letters · doi:10.1016/0009-2614(81)80303-x · Cited by 9
Local Disorder in Lithium Imide from Density Functional Simulation and NMR Spectroscopy
Bonnet, M., Iannuzzi, M., Sebastiani, D., & Hutter, J. (2012) · The Journal of Physical Chemistry C · doi:10.1021/jp3004272 · Cited by 9
Self-citationDynamical Dimension to the Hofmeister Series: Insights from First‐Principles Simulations
Bekçioğlu-Neff, G., Allolio, C., Desmukh, Y. S., Hansen, M. R., & Sebastiani, D. (2016) · ChemPhysChem · doi:10.1002/cphc.201501150 · Cited by 9
Using Approximate Computing for the Calculation of Inverse Matrix $p$ th Roots
Lass, M., Kühne, T. D., & Plessl, C. (2017) · IEEE Embedded Systems Letters · doi:10.1109/les.2017.2760923 · Cited by 8
Self-citationLuo, Z., Friščić, T., & Khaliullin, R. Z. (2017) · Physical Chemistry Chemical Physics · doi:10.1039/c7cp06828j · Cited by 8
Self-citationMüller, P., Karhan, K., Krack, M., Gerstmann, U., Schmidt, W. G., Bauer, M., & Kühne, T. D. (2018) · Journal of Computational Chemistry · doi:10.1002/jcc.25641 · Cited by 8
Self-citationAccurate Sampling with Noisy Forces from Approximate Computing
Rengaraj, V., Lass, M., Plessl, C., & Kühne, T. D. (2020) · Computation · doi:10.3390/computation8020039 · Cited by 8
Self-citationRichters, D., Lass, M., Walther, A., Plessl, C., & Kühne, T. D. (2018) · Communications in Computational Physics · doi:10.4208/cicp.oa-2018-0053 · Cited by 7
Self-citationSecond generation Car-Parrinello MD: application to the h-BN/Rh(111) nanomesh
Musso, T., Caravati, S., Hutter, J., & Iannuzzi, M. (2018) · The European Physical Journal B · doi:10.1140/epjb/e2018-90104-y · Cited by 7
Self-citationMicrocanonical RT-TDDFT simulations of realistically extended devices
Andermatt, S., Bani-Hashemian, M. H., Ducry, F., Brück, S., Clima, S., Pourtois, G., VandeVondele, J., & Luisier, M. (2018) · The Journal of Chemical Physics · doi:10.1063/1.5040048 · Cited by 6
Self-citationDiamondoid-functionalized nanogaps: from small molecules to electronic biosensing
Maier, F. C., Sarap, C. S., Dou, M., Sivaraman, G., & Fyta, M. (2019) · The European Physical Journal Special Topics · doi:10.1140/epjst/e2019-800157-0 · Cited by 6
From flat to tilted: gradual interfaces in organic thin film growth
Scarbath-Evers, L. K., Hammer, R., Golze, D., Brehm, M., Sebastiani, D., & Widdra, W. (2020) · Nanoscale · doi:10.1039/c9nr06592j · Cited by 6
Self-citationIdentifying Photoreaction Products in Cinnamate-Based Photoalignment Materials
Adams, D. J., Chappellet, S., Lincker, F., Ibn‐Elhaj, M., Watts, B., Iannuzzi, M., Šišak, D., Pignedoli, C. A., & Passerone, D. (2014) · The Journal of Physical Chemistry C · doi:10.1021/jp504765f · Cited by 5
Self-citationRichters, D., & Kühne, T. D. (2018) · Molecular Simulation · doi:10.1080/08927022.2018.1511899 · Cited by 5
Self-citationDirect Unconstrained Variable-Metric Localization of One-Electron Orbitals
Luo, Z., & Khaliullin, R. Z. (2020) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.9b01286 · Cited by 5
Self-citationPartovi‐Azar, P., Berg, M., Sanna, S., & Kühne, T. D. (2016) · International Journal of Quantum Chemistry · doi:10.1002/qua.25150 · Cited by 4
Self-citationInverse simulated annealing: Improvements and application to amorphous InSb
Los, J. H., Gabardi, S., Bernasconi, M., & Kühne, T. D. (2016) · Computational Materials Science · doi:10.1016/j.commatsci.2016.01.017 · Cited by 3
Self-citationLarge-Scale Nanoelectronic Device Simulation from First Principles
Bani-Hashemian, M. H. (2016) · Repository for Publications and Research Data (ETH Zurich) · doi:10.3929/ethz-a-010811338 · Cited by 3 · cited 2×
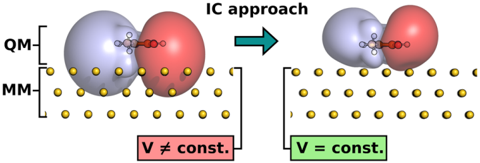
“composed of tens of thousands of atoms, the quantum transport solver OMEN has been integrated within CP2K [369–371].”
MethodSelf-citationRobust linear-scaling optimization of compact localized orbitals in density functional theoryPreprint
Shi, Y., Karaguesian, J., & Khaliullin, R. Z. (2020) · arXiv (Cornell University) · doi:10.48550/arxiv.2004.05901 · Cited by 3
Self-citation
Research works printed but not linked: 7
- Grotendorst, 2004, Quickstep: Make the atoms dance, 29
- 1969, ESAIM: Math. Modell. Numer. Anal., 3, 35
- 2006, Numerical Optimization
- B. Leimkuhler, M. Sachs, and G. Stoltz, “Hypocoercivity properties of adaptive Langevin dynamics,” arXiv:1908.09363 (2019). · arXiv · doi:10.1137/19m1291649
- Grotendorst, 2000, Modern Methods and Algorithms of Quantum Chemistry, 257
- 1964, Sov. Phys. JETP, 20, 1018
- 2014, Using Advanced MPI: Modern Features of the Message-Passing Interface
Cited by
A comprehensive electron wavefunction analysis toolbox for chemists, Multiwfn
Lu, T. (2024) · The Journal of Chemical Physics · doi:10.1063/5.0216272 · Cited by 5,213
Lu, T., & Chen, Q. (2022) · Journal of Computational Chemistry · doi:10.1002/jcc.26812 · Cited by 3,904
Lu, T., & Chen, Q. (2021) · Chemistry–Methods · doi:10.1002/cmtd.202100007 · Cited by 2,004
Extended tight‐binding quantum chemistry methods
Bannwarth, C., Caldeweyher, E., Ehlert, S., Hansen, A., Pracht, P., Seibert, J., Spicher, S., & Grimme, S. (2021) · WIREs Computational Molecular Science · doi:10.1002/wcms.1493 · Cited by 1,662
Shermo: A general code for calculating molecular thermochemistry properties
Lu, T., & Chen, Q. (2021) · Computational and Theoretical Chemistry · doi:10.1016/j.comptc.2021.113249 · Cited by 1,402
Metadata
- Authors
- Kühne, T. D., Iannuzzi, M., Del Ben, M., Rybkin, V. V., Seewald, P., Stein, F., Laino, T., Khaliullin, R. Z., Schütt, O., Schiffmann, F., Golze, D., Wilhelm, J., Chulkov, S., Bani-Hashemian, M. H., Weber, V., Borštnik, U., Taillefumier, M., Jakobovits, A. S., Lazzaro, A., Pabst, H., Müller, T., Schade, R., Guidon, M., Andermatt, S., Holmberg, N., Schenter, G. K., Hehn, A., Bussy, A., Belleflamme, F., Tabacchi, G., Glöß, A., Lass, M., Bethune, I., Mundy, C. J., Plessl, C., Watkins, M., VandeVondele, J., Krack, M., & Hutter, J.
- Affiliations
- Paderborn University (DE)
- University of Zurich (CH)
- Computational Research Division, Lawrence Berkeley National Laboratory 3 , Berkeley, California 94720, USA
- IBM Research - Zurich (CH)
- McGill University (CA)
- ETH Zurich (CH)
- Victoria University (AU)
- Aalto University (FI)
- University of Regensburg (DE)
- University of Lincoln (GB)
- HPE Switzerland GmbH 14 , Basel, Switzerland
- Intel (United States) (US)
- Pacific Northwest National Laboratory (US)
- University of Insubria (IT)
- BASF (Germany) (DE)
- Science and Technology Facilities Council (GB)
- School of Mathematics and Physics, University of Lincoln 10 , Brayford Pool, Lincoln, United Kingdom
- Paul Scherrer Institute (CH)
- Corresponding author
- Kühne, Thomas D.
- ORCID
- Kühne, Thomas D.
- Iannuzzi, Marcella
- Rybkin, Vladimir V.
- Stein, Frederick
- Laino, Teodoro
- Khaliullin, Rustam Z.
- Schütt, Ole
- Schiffmann, Florian
- Golze, Dorothea
- Wilhelm, Jan
- Chulkov, Sergey
- Bani-Hashemian, Mohammad Hossein
- Taillefumier, Mathieu
- Jakobovits, Alice Shoshana
- Lazzaro, Alfio
- Müller, Tiziano
- Schade, Robert
- Guidon, Manuel
- Andermatt, Samuel
- Holmberg, Nico
- Schenter, Gregory K.
- Hehn, Anna
- Bussy, Augustin
- Belleflamme, Fabian
- Tabacchi, Gloria
- Lass, Michael
- Bethune, Iain
- Mundy, Christopher J.
- Plessl, Christian
- Watkins, Matt
- VandeVondele, Joost
- Krack, Matthias
- Hutter, Jürg
- Venue
- The Journal of Chemical Physics
- Publisher
- AIP Publishing
- Published
- 2020-05-19
- Volume
- 152
- Issue
- 19
- Pages
- 194103-194103
- ISSN
- 0021-9606 (ISSN-L)
- 1089-7690
- OpenAlex
- W3027308092
- Cited by
- 4,468
- References
- 445
- Citations by year
- 2026: 1066 · 2025: 1195 · 2024: 870 · 2023: 561 · 2022: 431 · 2021: 284 · 2020: 58
- FWCI
- 135.74
- Type
- journal article
- Language
- en
- Open access
- hybrid
- doi.org
- Licence
- cc-by
- creativecommons.org/licenses/by/4.0/ (vor)
- creativecommons.org/licenses/by/4.0/ (tdm)
- Locations
- The Journal of Chemical Physics (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Repository for Publications and Research Data (ETH Zurich) (publishedVersion, cc-by)
- Lincoln Repository (University of Lincoln) (acceptedVersion)
- University of Regensburg Publication Server (University of Regensburg) (acceptedVersion)
- Zurich Open Repository and Archive (University of Zurich) (acceptedVersion, cc-by)
- arXiv (Cornell University) (submittedVersion)
- Victoria University Research Repository (Victoria University) (submittedVersion, cc-by)
- Aaltodoc (Aalto University) (submittedVersion, cc-by)
- DORA PSI (Paul Scherrer Institute) (submittedVersion, cc-by)
- Figshare (submittedVersion, cc-by)
- IrInSubria (University of Insubria) (submittedVersion, other-oa)
- OSTI OAI (U.S. Department of Energy Office of Scientific and Technical Information) (submittedVersion)
- Zurich Open Repository and Archive (University of Zurich) (submittedVersion, cc-by)
- arXiv (Cornell University)
- ETH Zürich Research Collection (cc-by)
- Zurich Open Repository and Archive (University of Zurich)
- University of Regensburg Publication Server (University of Regensburg)
- Funders
- Natural Sciences and Engineering Research Council of Canada (RGPIN-2016-0505)
- H2020 European Research Council (716142)
- Schweizerischer Nationalfonds Zur Förderung der Wissenschaftlichen Forschung (PZ00P2 174227)
- Topics
- Physical Sciences › Physics and Astronomy › Atomic and Molecular Physics, and Optics › Advanced Chemical Physics Studies
- Physical Sciences › Chemistry › Physical and Theoretical Chemistry › Advanced Physical and Chemical Molecular Interactions
- Physical Sciences › Computer Science › Hardware and Architecture › Parallel Computing and Optimization Techniques
- Keywords
- CP2K, electronic structure software, molecular dynamics, ab initio molecular dynamics, density functional theory, post-Hartree-Fock methods, Gaussian basis sets, plane-wave basis, linear-scaling methods, massively parallel computing, atomistic simulation
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
Data sources
Bibliographic data: Semantic Scholar, OpenAlex, Crossref.
Contains information from the Semantic Scholar Academic Graph Datasets, made available under the ODC Attribution License (ODC-BY 1.0).
Semantic Scholar’s data is described in: Kinney, R., et al. (2023). The Semantic Scholar Open Data Platform. arXiv:2301.10140.
