

The DOI matches its Crossref record; 2 of the 2 references it cites were found in a registry or an index.
Deciphering GB1’s Single Mutational Landscape: Insights from MuMi Analysis
Open accessPreprint — published as 10.1021/acs.jpcb.4c04916 · Paper page
Abstract
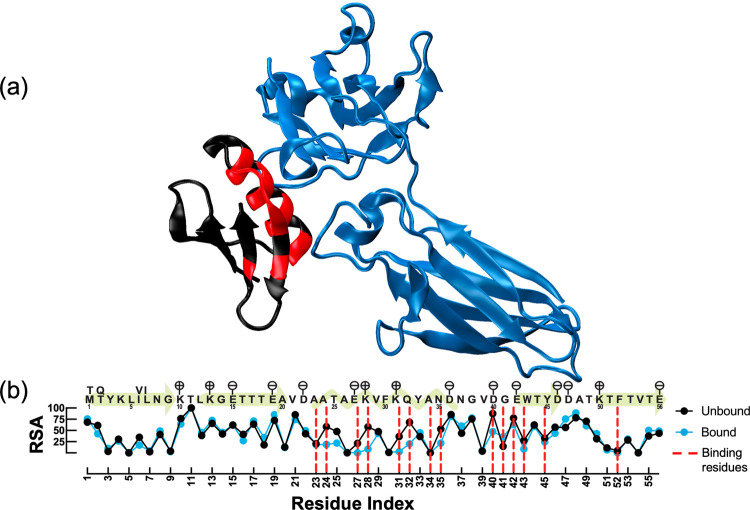
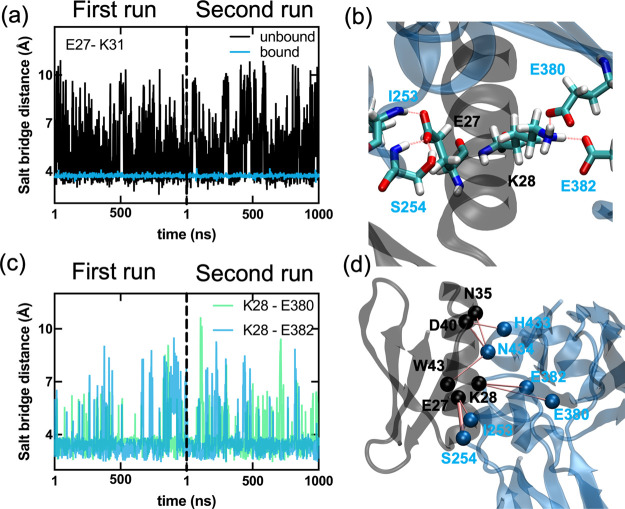
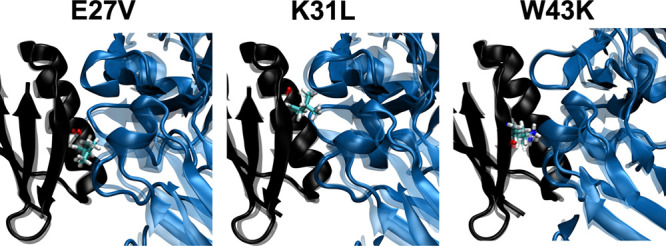
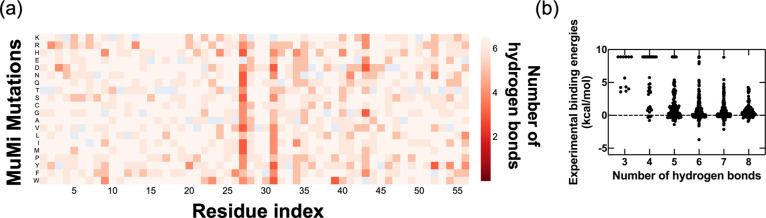
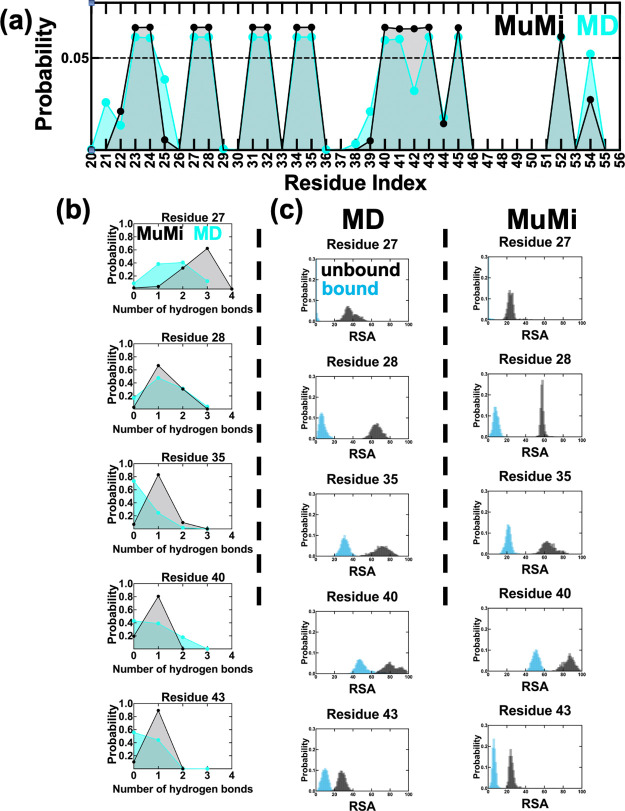
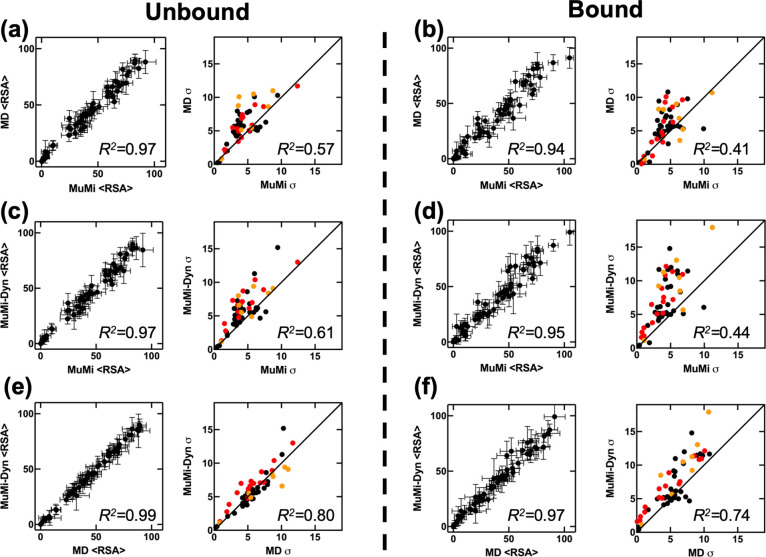
Mutational changes that affect the binding of the C2 fragment of Streptococcal protein G (GB1) to the Fc domain of human IgG (IgG-Fc) have been extensively studied using deep mutational scanning (DMS), and the binding affinity of all single mutations has been measured experimentally in the literature. To investigate the underlying molecular basis, we perform in-silico mutational scanning for all possible single mutations, along with 2-µs-long molecular dynamics (WT-MD) of the wild-type (WT) GB1 in both unbound and IgG-Fc bound forms. We compute the hydrogen bonds between GB1 and IgG-Fc in WT-MD to identify the dominant hydrogen bonds for binding, which we then assess in conformations produced by Mutation and Minimization (MuMi) to explain the fitness landscape of GB1 and IgG-Fc binding. Furthermore, we analyze MuMi and WT-MD to investigate the dynamics of binding, focusing on the relative solvent accessibility (RSA) of residues and the probability of residues being located at the binding interface. With these analyses, we explain the interactions between GB1 and IgG-Fc and display the structural features of binding. Our findings pave the way for improved predictive accuracy in protein stability and interaction studies, which are crucial for advancements in drug design and synthetic biology.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Highly accurate protein structure prediction with AlphaFold
Jumper, J., Evans, R., Pritzel, A., Green, T., Figurnov, M., Ronneberger, O., Tunyasuvunakool, K., Bates, R., Žídek, A., Potapenko, A., et al. (2021) · Nature · doi:10.1038/s41586-021-03819-2 · Cited by 43,764
Sampling alternative conformational states of transporters and receptors with AlphaFold2
del Alamo, D., Sala, D., Mchaourab, H. S., & Meiler, J. (2022) · eLife · doi:10.7554/elife.75751 · Cited by 511
Research works printed but not linked: 40
- 2014, Deep mutational scanning: a new style of protein science, Nature Methods, 11, 801, 10.1038/nmeth.3027 · DOI · Paper page
- 10.1128/JB.00070-11 · DOI
- 10.1016/j.tibtech.2011.04.003 · DOI
- 2023, Accurate proteome-wide missense variant effect prediction with AlphaMissense, Science, 381, 1303
- Diaz, D. J. , Kulikova, A. V. , Ellington, A. D. & Wilke, C. O . Using machine learning to predict the effects and consequences of mutations in proteins. Current Opinion in Structural Biology 78, (2023). · DOI
- 2023, Discovering functionally important sites in proteins, Nature Communications, 14, 4175, 10.1038/s41467-023-39909-0 · DOI
- 2024, ProSTAGE: Predicting Effects of Mutations on Protein Stability by Using Protein Embeddings and Graph Convolutional Networks, Journal of Chemical Information and Modeling, 64, 340, 10.1021/acs.jcim.3c01697 · DOI
- Blaabjerg, L. M. et al. Rapid protein stability prediction using deep learning representations. eLife 12, (2023). · DOI · Paper page
- 2018, Biophysical inference of epistasis and the effects of mutations on protein stability and function, Molecular Biology and Evolution, 35, 2345, 10.1093/molbev/msy141 · DOI
- 10.1016/j.jmb.2016.12.007 · DOI
- 10.1101/2024.03.15.585128 · DOI
- 10.1038/s41586-022-04586-4 · DOI
- Dewachter, L. , Brooks, F. D. , Wolff, M. H. , White, A. R. & Moore, M. J . Deep mutational scanning of essential bacterial proteins can guide antibiotic development. Nature Communications 14, (2023). · DOI · Paper page
- 2021, Climbing Up and Down Binding Landscapes through Deep Mutational Scanning of Three Homologous Protein–Protein Complexes, J. Am. Chem. Soc, 143, 17261, 10.1021/jacs.1c08707 · DOI
- 2021, Structurally Distributed Surface Sites Tune Allosteric Regulation, Protein Science, 30, 133
- 10.1016/j.cub.2014.09.072 · DOI
- Wu, N. C. , Dai, L. , Olson, C. A. , Lloyd-Smith, J. O. & Sun, R . Adaptation in protein fitness landscapes is facilitated by indirect paths. eLife 5, (2016). · DOI
- 10.1073/pnas.1903888116 · DOI
- 2023, Mega-scale experimental analysis of protein folding stability in biology and design, Nature, 620, 434, 10.1038/s41586-023-06328-6 · DOI · Paper page
- 10.1038/s41588-019-0431-x · DOI
- 10.1038/s41588-019-0432-9 · DOI
- 10.1016/j.sbi.2021.11.001 · DOI
- 2001, Elucidating the structural mechanisms for biological activity of the chemokine family, Proteins: Structure, Function, and Bioinformatics, 43, 150, 10.1002/1097-0134(20010501)43:2<150::AID-PROT1027>3.0.CO;2-M · DOI
- 1996, New conformational search method based on local torsional deformations for cyclic molecules, loops in proteins, and dense polymer systems, Journal of Chemical Physics, 105, 7868, 10.1063/1.472607 · DOI
- 1997, Efficiency of the local torsional deformations method for identifying the stable structures of cyclic molecules, Journal of Physical Chemistry A, 101, 2185, 10.1021/jp9627537 · DOI
- 2015, In silico mutational studies of Hsp70 disclose sites with distinct functional attributes, Proteins: Structure, Function and Bioinformatics, 83, 2077, 10.1002/prot.24925 · DOI
- 10.1038/S41592-022-01488-1 · DOI · Paper page
- 10.1093/nar/28.1.235 · DOI · Paper page
- 10.1016/0263-7855(96)00018-5 · DOI · Paper page
- 2021, ProDy 2.0: increased scale and scope after 10 years of protein dynamics modelling with Python, Bioinformatics, 37, 3657, 10.1093/bioinformatics/btab187 · DOI
- 1999, NAMD2: greater scalability for parallel molecular dynamics, J Comput Phys, 151, 283, 10.1006/jcph.1999.6201 · DOI
- 2012, Optimization of the Additive CHARMM All-Atom Protein Force Field Targeting Improved Sampling of the Backbone ϕ, ψ and Side-Chain χ1 and χ2 Dihedral Angles, Journal of Chemical Theory and Computation, 8, 3257, 10.1021/ct300400x · DOI
- 10.1038/nbt.3988 · DOI · Paper page
- Mitternacht, S . FreeSASA: An open source C library for solvent accessible surface area calculations [version 1; peer review: 2 approved]. F1000Research vol. 5 (2016). · DOI
- 2016, High-throughput identification of protein mutant stability computed from a double mutant fitness landscape, Protein Science, 25, 530, 10.1002/pro.2840 · DOI
- 10.1002/prot.1124 · DOI
- 10.1002/prot.26688 · DOI · Paper page
- 2022, Ligand binding remodels protein side-chain conformational heterogeneity, Elife, 11, e74114, 10.7554/eLife.74114 · DOI
- 2021, N-Terminus of the Third PDZ Domain of PSD-95 Orchestrates Allosteric Communication for Selective Ligand Binding, Journal of Chemical Information and Modeling, 61, 347, 10.1021/acs.jcim.0c01079 · DOI
- 2021, Dynamic Community Composition Unravels Allosteric Communication in PDZ3, Journal of Physical Chemistry B, 125, 2266, 10.1021/acs.jpcb.0c11604 · DOI
Metadata
- Authors
- Guclu, T. F., Atilgan, A. R., & Atilgan, C.
- Affiliations
- Sabancı Üniversitesi (TR)
- Corresponding author
- Guclu, Tandac F.
- Venue
- The Journal of Physical Chemistry B
- Publisher
- openRxiv
- Published
- 2024-06-03
- Volume
- 128
- Issue
- 33
- Pages
- 7987-7996
- ISSN
- 1520-5207 (ISSN-L)
- PMC
- PMC11671028
- OpenAlex
- W4401425370
- Cited by
- 0
- References
- 2
- FWCI
- 0.00
- Type
- journal article
- Language
- en
- Open access
- hybrid
- doi.org
- Licence
- cc-by
- Locations
- The Journal of Physical Chemistry B (publishedVersion, cc-by)
- bioRxiv (Cold Spring Harbor Laboratory) (acceptedVersion, cc-by-nc)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Sabanci University (acceptedVersion)
- Sabanci University (submittedVersion)
- Funders
- Türkiye Bilimsel ve Teknolojik Araştırma Kurumu (122F149, 121Z329)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Protein Structure and Dynamics
- Life Sciences › Immunology and Microbiology › Immunology › Monoclonal and Polyclonal Antibodies Research
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › vaccines and immunoinformatics approaches
- Keywords
- streptococcal protein G, GB1, IgG Fc binding, deep mutational scanning, fitness landscape, hydrogen bonding, protein-protein binding, solvent accessibility, binding interface, protein stability, molecular dynamics
- MeSH terms
- Bacterial Proteins, Humans, Hydrogen Bonding, Immunoglobulin G, Immunoglobulin Fc Fragments, Mutation, Protein Binding, Molecular Dynamics Simulation
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
