

The DOI matches its Crossref record; 1 of the 1 references it cites were found in a registry or an index.
Protocols for Multi-Scale Molecular Dynamics Simulations: A Comparative Study for Intrinsically Disordered Amyloid Beta in Amber & Gromacs on CPU & GPU
Open accessPreprint — published as 10.3390/ijms25126698
Abstract
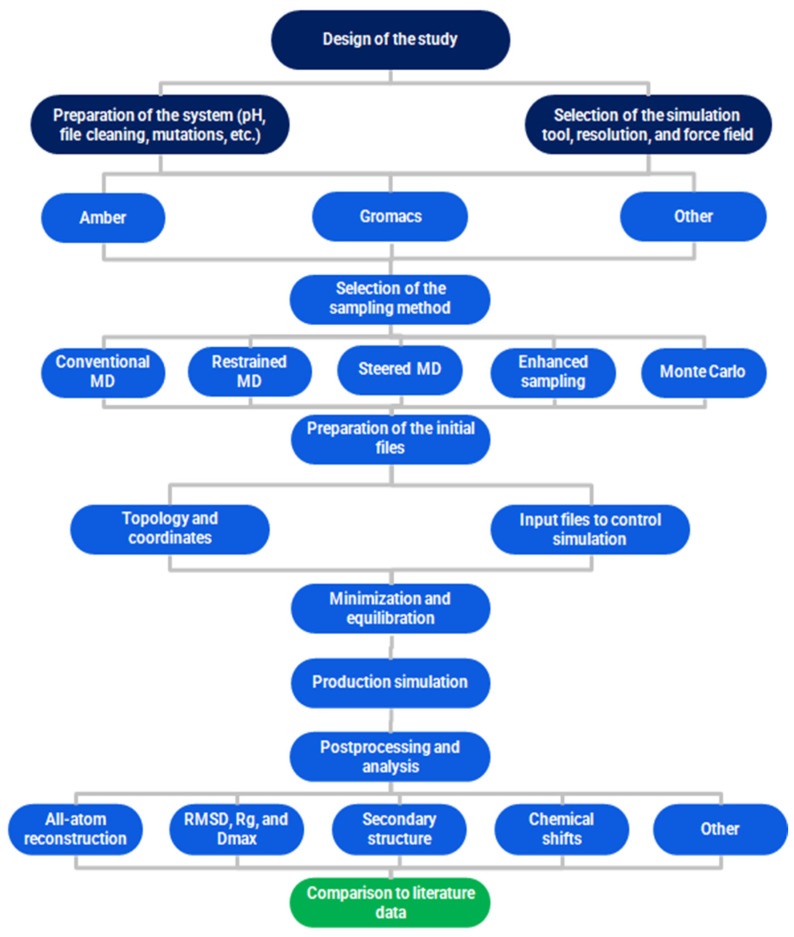
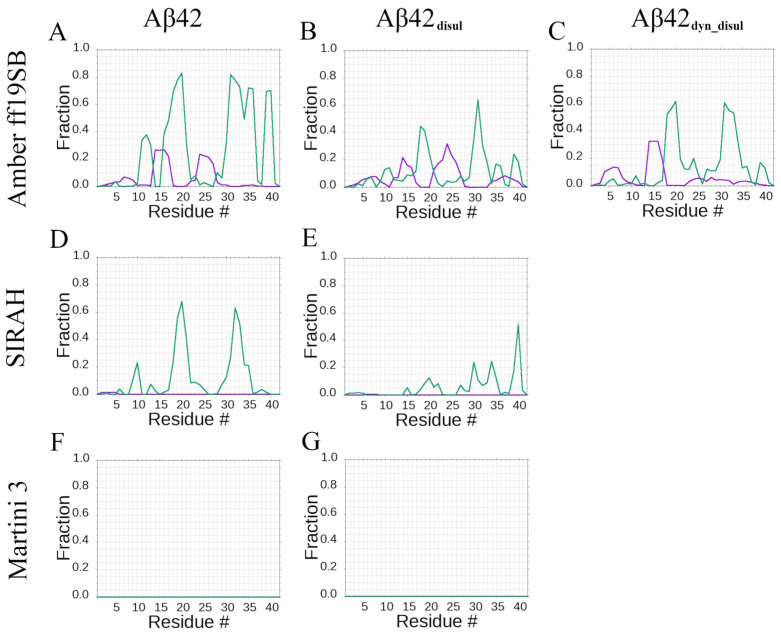
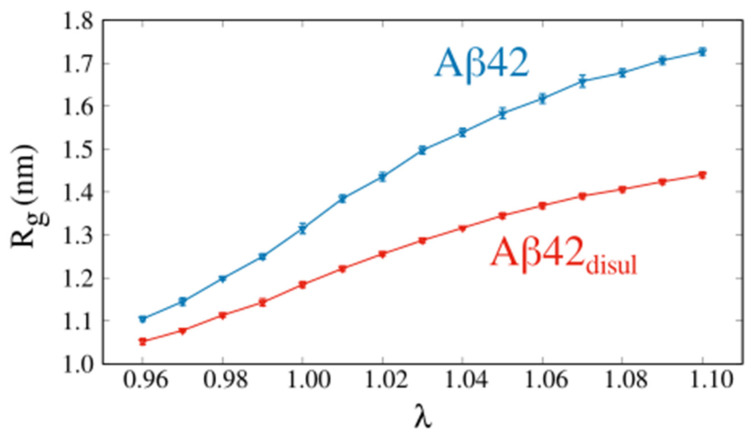
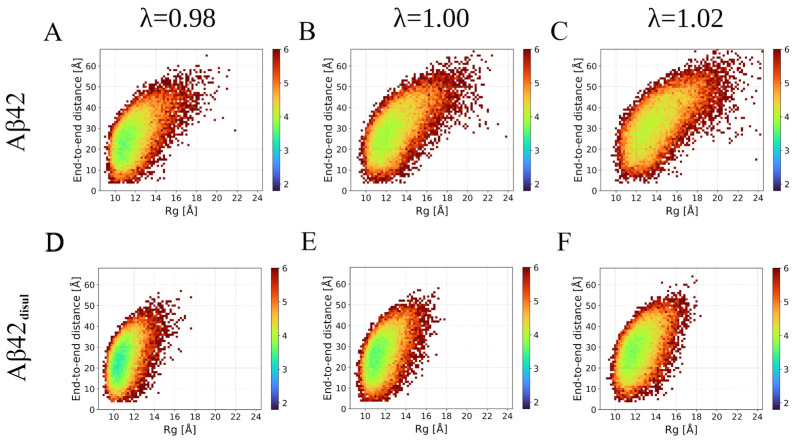
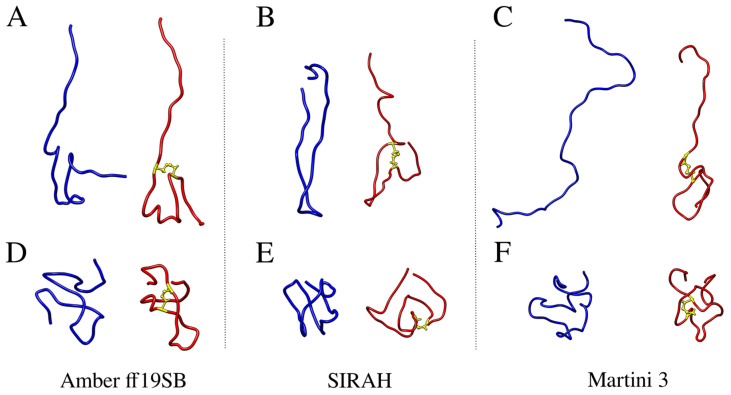
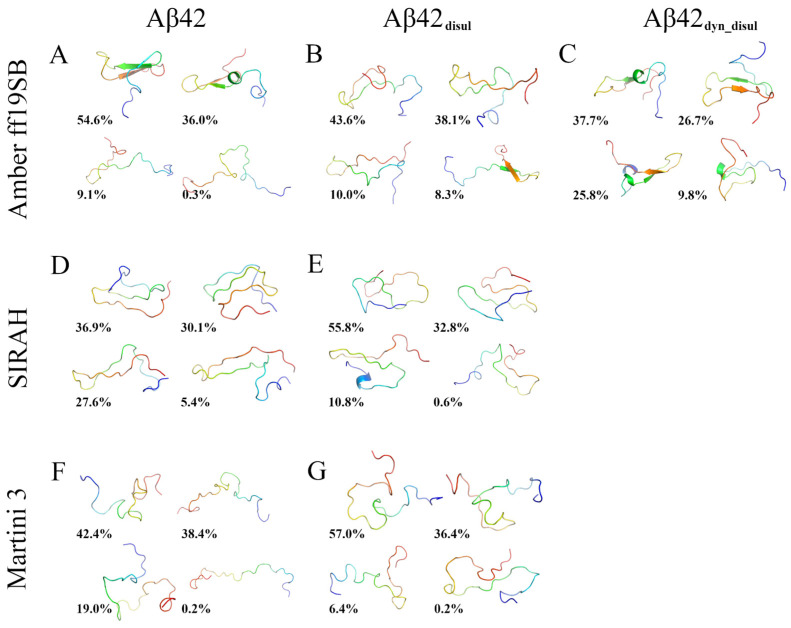
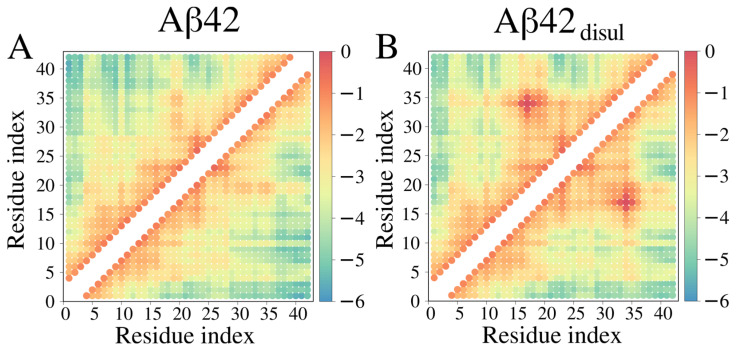
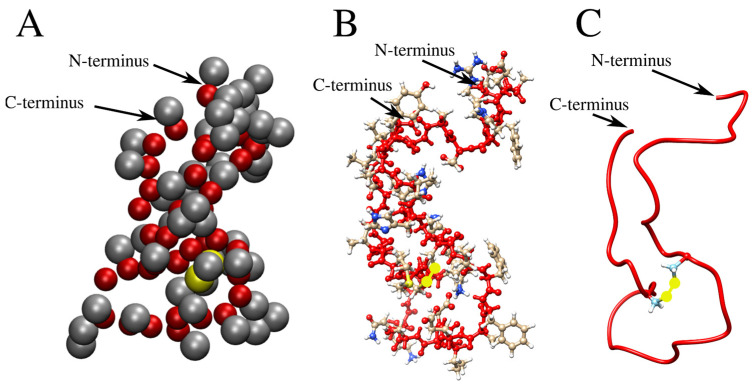
Intrinsically disordered proteins (IDPs) present challenges to conventional experimental techniques due to their large-scale conformational fluctuations and the transient occurrence of structural elements. This work illustrates computational methods for studying IDPs at various levels of resolution. The included simulation protocol offers a step-by-step guide on how to conduct molecular dynamics (MD) simulations and analyze the results using the Amber and Gromacs packages, employing both all-atom and coarse-grained approaches. This protocol can be easily adapted to study other biomacromolecules, including folded and disordered proteins and peptides. Furthermore, it is discussed in this work how to perform standard molecular modeling operations, such as amino-acid substitutions (mutagenesis) and insertions of residues missing in a protein structure, as well as how to incorporate post-translational modifications into the simulations, such as disulfide bonds, which are often crucial for proteins to attain their physiologically functional structure. In conventional MD studies, disulfide bonds are typically fixed at the preparation step and remain unchanged throughout the simulations, unable to break or reform. Here, in contrast, a dynamic approach is presented. It involves adequate distance restraints applied to the sulfur atoms of selected cysteine residues, allowing disulfide bonds to break and reform during the simulation. The effectiveness of these methodologies is demonstrated by examining a model IDP, the monomeric form of 1-42 amyloid-β (Aβ42), both with and without disulfide bonds, at different levels of resolution. This study not only contributes to our understanding of the role of disulfide bonds but also provides detailed simulation protocols that can serve as a foundation for future investigations. SUMMARY: Given the challenges of experimental studies on intrinsically disordered proteins, this manuscript demonstrates step-by-step protocols for conducting all-atom and coarse-grained molecular dynamics simulations using two widespread packages, Amber and Gromacs. The monomeric form of 1-42 amyloid-β (Aβ42) is used as an example, from which insights into the structure, dynamics and physicochemical properties of this protein can be obtained.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
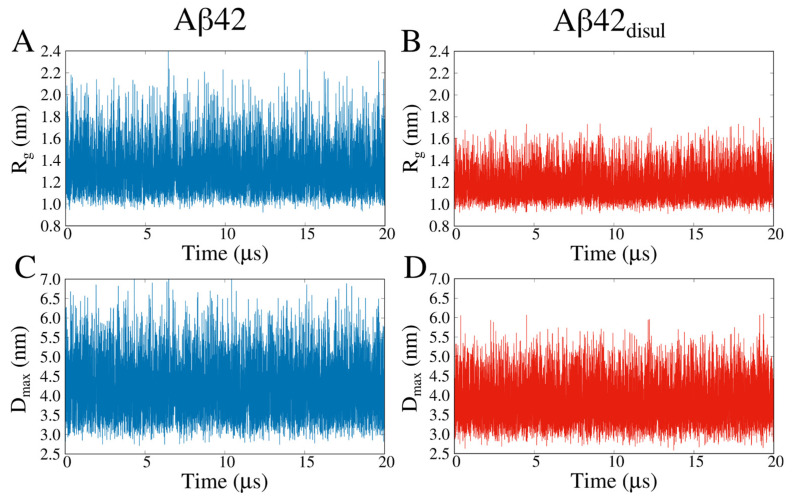
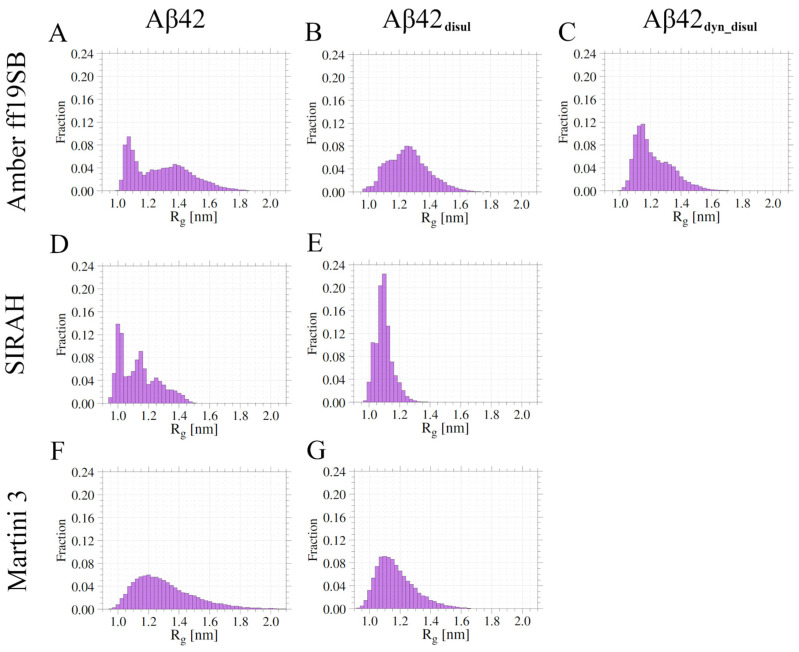
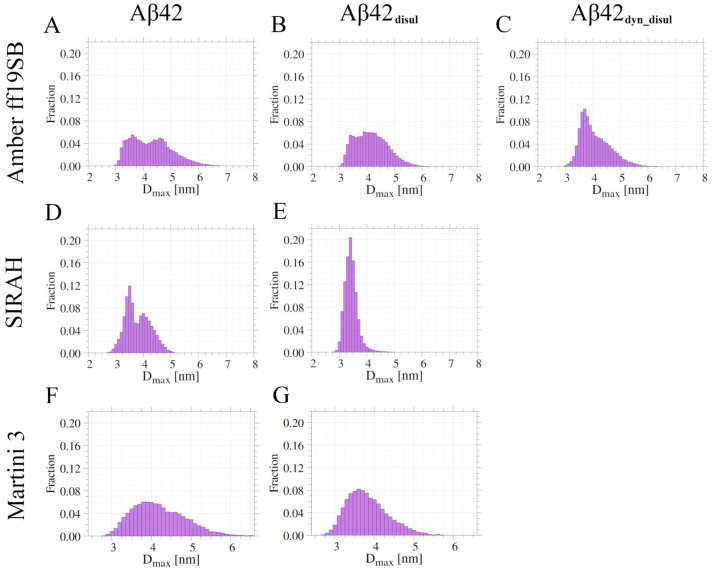
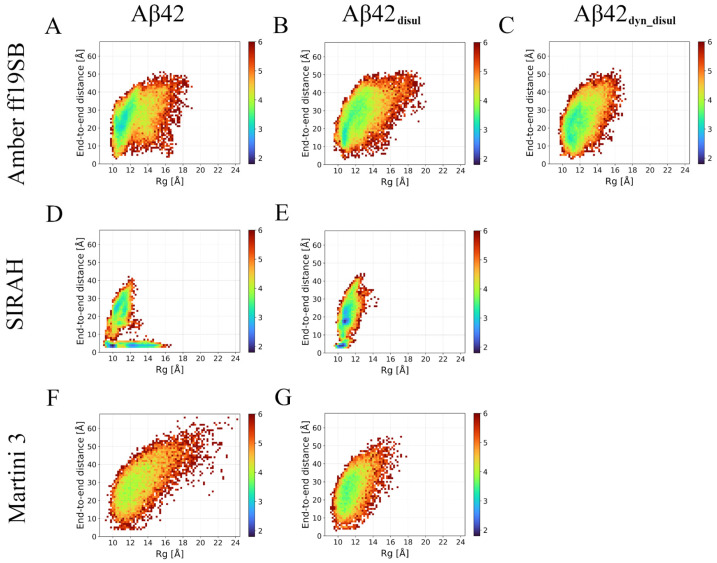
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Highly accurate protein structure prediction with AlphaFold
Jumper, J., Evans, R., Pritzel, A., Green, T., Figurnov, M., Ronneberger, O., Tunyasuvunakool, K., Bates, R., Žídek, A., Potapenko, A., et al. (2021) · Nature · doi:10.1038/s41586-021-03819-2 · Cited by 43,764
Research works printed but not linked: 63
- Whitford, D. Proteins: Structure and Function. Wiley. (2005).
- 10.1126/science.181.4096.223 · DOI · Paper page
- 10.1021/bi012159+ · DOI · Paper page
- 10.1016/j.jmb.2007.02.093 · DOI
- 10.1007/s10930-009-9201-4 · DOI
- 10.1073/pnas.1418155112 · DOI
- 2016, The Role of Amyloid-β Oligomers in Toxicity, Propagation, and Immunotherapy, EBioMedicine., 6, 42, 10.1016/j.ebiom.2016.03.035 · DOI · Paper page
- Breijyeh, Z. , Karaman, R. Comprehensive Review on Alzheimer’s Disease: Causes and Treatment. Molecules. 25 (24) (2020). · DOI · Paper page
- 10.1021/bi048019s · DOI
- 2020, Cysteines and Disulfide Bonds as Structure-Forming Units: Insights From Different Domains of Life and the Potential for Characterization by NMR, Frontiers in chemistry., 8, 280, 10.3389/fchem.2020.00280 · DOI
- 10.1016/j.biotechadv.2004.09.002 · DOI
- 2022, Mechanical Stability of Ribonuclease A Heavily Depends on the Redox Environment, The journal of physical chemistry. B., 126, 6240, 10.1021/acs.jpcb.2c04718 · DOI
- 10.1063/1.5093184 · DOI
- 2021, Recent Force Field Strategies for Intrinsically Disordered Proteins, Journal of chemical information and modeling., 61, 1037, 10.1021/acs.jcim.0c01175 · DOI
- 2021, Assessing SIRAH’s Capability to Simulate Intrinsically Disordered Proteins and Peptides, Journal of chemical theory and computation., 17, 599, 10.1021/acs.jctc.0c00948 · DOI
- 10.1002/prot.21123 · DOI
- 2015, ff14SB: Improving the Accuracy of Protein Side Chain and Backbone Parameters from ff99SB, Journal of chemical theory and computation., 11, 3696, 10.1021/acs.jctc.5b00255 · DOI
- 10.1002/jcc.20065 · DOI
- 10.1002/jcc.23354 · DOI
- 2020, ff19SB: Amino-Acid-Specific Protein Backbone Parameters Trained against Quantum Mechanics Energy Surfaces in Solution, Journal of chemical theory and computation., 16, 528, 10.1021/acs.jctc.9b00591 · DOI
- 2017, CHARMM36m: an improved force field for folded and intrinsically disordered proteins, Nature methods., 14, 71, 10.1038/nmeth.4067 · DOI
- 2015, Water dispersion interactions strongly influence simulated structural properties of disordered protein states, The journal of physical chemistry. B., 119, 5113, 10.1021/jp508971m · DOI
- 10.1021/acs.jcim.0c01390 · DOI
- 2015, The Dependence of Amyloid-β Dynamics on Protein Force Fields and Water Models, Chemphyschem: a European journal of chemical physics and physical chemistry., 16, 3278, 10.1002/cphc.201500415 · DOI
- 10.1016/S0009-2614(99)01123-9 · DOI
- Qi, R. , Wei, G. , Ma, B. , Nussinov, R. Replica Exchange Molecular Dynamics: A Practical Application Protocol with Solutions to Common Problems and a Peptide Aggregation and Self-Assembly Example. Peptide Self-Assembly: Methods and Protocols. 101–119 (2018). · DOI
- 10.1021/acs.jcim.8b00462 · DOI
- 10.1021/acs.chemrev.6b00163 · DOI
- 2022, Unifying coarse-grained force fields for folded and disordered proteins, Current opinion in structural biology., 72, 63, 10.1016/j.sbi.2021.08.006 · DOI
- 2014, A unified coarse-grained model of biological macromolecules based on mean-field multipole-multipole interactions, Journal of molecular modeling., 20, 2306, 10.1007/s00894-014-2306-5 · DOI
- 2020, Chapter Two - Scale-consistent approach to the derivation of coarse-grained force fields for simulating structure, dynamics, and thermodynamics of biopolymers, Progress in Molecular Biology and Translational Science., 170, 73, 10.1016/bs.pmbts.2019.12.004 · DOI
- 2018, AWSEM-IDP: A coarse-grained force field for intrinsically disordered proteins, The journal of physical chemistry. B., 122, 11115, 10.1021/acs.jpcb.8b05791 · DOI
- 2021, Martini 3: a general purpose force field for coarse-grained molecular dynamics, Nature methods., 18, 382, 10.1038/s41592-021-01098-3 · DOI · Paper page
- 2022, Improving Martini 3 for Disordered and Multidomain Proteins, Journal of chemical theory and computation., 18, 2033, 10.1021/acs.jctc.1c01042 · DOI
- 2023, The SIRAH force field: A suite for simulations of complex biological systems at the coarse-grained and multiscale levels, Journal of structural biology., 215, 107985, 10.1016/j.jsb.2023.107985 · DOI
- Bekker, H. et al. GROMACS - A Parallel Computer For Molecular-Dynamics Simulations. 4th International Conference on Computational Physics (PC 92). 252–256 (1993).
- 2017, Dynamics of Disulfide-Bond Disruption and Formation in the Thermal Unfolding of Ribonuclease A, Journal of chemical theory and computation., 13, 5721, 10.1021/acs.jctc.7b00724 · DOI
- 2018, Improvements to the APBS biomolecular solvation software suite, Protein science: a publication of the Protein Society., 27, 112, 10.1002/pro.3280 · DOI
- 10.1002/jcc.20945 · DOI · Paper page
- 2016, Comparative Protein Structure Modeling Using MODELLER, Current protocols in bioinformatics., 54, 5.6.1
- 2015, Protein Structure and Function Prediction Using I-TASSER, Current protocols in bioinformatics., 52, 5.8.1
- Vanquelef, E. et al. R.E.D. Server: a web service for deriving RESP and ESP charges and building force field libraries for new molecules and molecular fragments. Nucleic acids research. 39 (Web Server issue), W511–7 (2011). · DOI
- 2017, ff14IDPs force field improving the conformation sampling of intrinsically disordered proteins, Chemical biology & drug design., 89, 5, 10.1111/cbdd.12832 · DOI
- Yang, J. , Zhang, Y. I-TASSER server: new development for protein structure and function predictions. Nucleic acids research. 43 (W1), W174–81 (2015). · DOI
- 2014, Going Backward: A Flexible Geometric Approach to Reverse Transformation from Coarse Grained to Atomistic Models, Journal of chemical theory and computation., 10, 676, 10.1021/ct400617g · DOI
- 10.1002/(SICI)1096-987X(199709)18:12<1463::AID-JCC4>3.3.CO;2-L · DOI
- 2009, Combining an Elastic Network With a Coarse-Grained Molecular Force Field: Structure, Dynamics, and Intermolecular Recognition, Journal of chemical theory and computation., 5, 2531, 10.1021/ct9002114 · DOI
- 2016, Improved Side Chain Dynamics in MARTINI Simulations of Protein-Lipid Interfaces, Journal of chemical theory and computation., 12, 2446, 10.1021/acs.jctc.6b00122 · DOI
- 2014, Balanced Protein-Water Interactions Improve Properties of Disordered Proteins and Non-Specific Protein Association, Journal of chemical theory and computation., 10, 5113, 10.1021/ct500569b · DOI
- 2014, Building Water Models: A Different Approach, Journal of Physical Chemistry Letters., 5, 3863, 10.1021/jz501780a · DOI
- 2011, Physiologically relevant concentrations of NaCl and KCl increase DNA photocleavage by an N-substituted 9-aminomethylanthracene dye, Biochemistry., 50, 10375, 10.1021/bi200972c · DOI
- 2022, Conformational ensemble of the full-length SARS-CoV-2 nucleocapsid (N) protein based on molecular simulations and SAXS data, Biophysical chemistry., 288, 106843, 10.1016/j.bpc.2022.106843 · DOI
- Dignon, G.L. , Zheng, W. , Kim, Y.C. , Best, R.B. , Mittal, J. Sequence determinants of protein phase behavior from a coarse-grained model. PLoS computational biology. 14 (1) (2018). · DOI
- 2023, Membrane curvature sensing by model biomolecular condensates, Soft matter., 19, 3723, 10.1039/D3SM00131H · DOI
- 2022, Modeling structural interconversion in Alzheimers’ amyloid beta peptide with classical and intrinsically disordered protein force fields, Journal of biomolecular structure & dynamics., 40, 10005, 10.1080/07391102.2021.1939163 · DOI
- 10.1016/S0006-3495(01)75993-0 · DOI
- 10.1385/CBB:46:2:123 · DOI
- 10.1074/jbc.M110.199885 · DOI
- Festa, G. et al. Aggregation States of Aβ1-40, Aβ1-42 and Aβp3-42 Amyloid Beta Peptides: A SANS Study. International journal of molecular sciences. 20 (17) (2019). · DOI
- 2016, Monomeric Amyloid Beta Peptide in Hexafluoroisopropanol Detected by Small Angle Neutron Scattering, PloS one., 11, e0150267, 10.1371/journal.pone.0150267 · DOI
- 10.1073/pnas.1518298112 · DOI
- 10.1002/jcc.20303 · DOI
- 10.1002/jcc.20035 · DOI
Metadata
- Authors
- Smardz, P., Anila, M. M., Rogowski, P., Li, M. S., Różycki, B., & Krupa, P.
- Affiliations
- Institute of Physics (PL)
- Venue
- International Journal of Molecular Sciences
- Publisher
- openRxiv
- Published
- 2023-10-24
- Volume
- 25
- Issue
- 12
- Pages
- 6698-6698
- ISSN
- 1422-0067 (ISSN-L)
- PMC
- PMC11204378
- OpenAlex
- W4399863716
- Cited by
- 26
- References
- 1
- Citations by year
- 2026: 10 · 2025: 12 · 2024: 4
- FWCI
- 3.53
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- cc-by
- Locations
- International Journal of Molecular Sciences (publishedVersion, cc-by)
- bioRxiv (Cold Spring Harbor Laboratory) (acceptedVersion, cc-by-nc-nd)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Funders
- Narodowym Centrum Nauki (2020/39/B/NZ1/00377, 2019/35/D/ST4/03156)
- China Scholarship Council
- Infrastruktura PL-Grid
- Academic Computer Centre Cyfronet, AGH University of Science and Technology
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Protein Structure and Dynamics
- Life Sciences › Neuroscience › Cellular and Molecular Neuroscience › Alzheimer's disease research and treatments
- Physical Sciences › Chemistry › Physical and Theoretical Chemistry › Advanced Physical and Chemical Molecular Interactions
- Keywords
- intrinsically disordered proteins, amyloid-beta 42, disulfide bonds, amyloid fibril formation, all-atom molecular dynamics, coarse-grained molecular dynamics, Martini 3 force field, biography of Muhammad, OPC water model, secondary structure content, chemical shift prediction, protein aggregation
- MeSH terms
- Disulfides, Humans, Peptide Fragments, Protein Conformation, Amyloid beta-Peptides, Protein Structure, Secondary, Molecular Dynamics Simulation, Intrinsically Disordered Proteins
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
How to cite
Cite CoChat for the checks on this page, and the paper for its own findings.
- This page
- CoChat. (2026). Citation check: Protocols for Multi-Scale Molecular Dynamics Simulations: A Comparative Study for Intrinsically Disordered Amyloid Beta in Amber & Gromacs on CPU & GPU. https://app.cochat.ai/papers/10.1101/2023.10.24.563575
- The paper
- Smardz, P., Anila, M. M., Rogowski, P., Li, M. S., Różycki, B., & Krupa, P. (2023). Protocols for Multi-Scale Molecular Dynamics Simulations: A Comparative Study for Intrinsically Disordered Amyloid Beta in Amber & Gromacs on CPU & GPU. International Journal of Molecular Sciences. https://doi.org/10.1101/2023.10.24.563575
