

The DOI matches its Crossref record; 40 of the 40 references it cites were found in a registry or an index.
Clinical and molecular characterization of 40 patients with classic Ehlers-Danlos syndrome: Identification of 18 COL5A1 and 2 COL5A2 novel mutations
Open accessAbstract
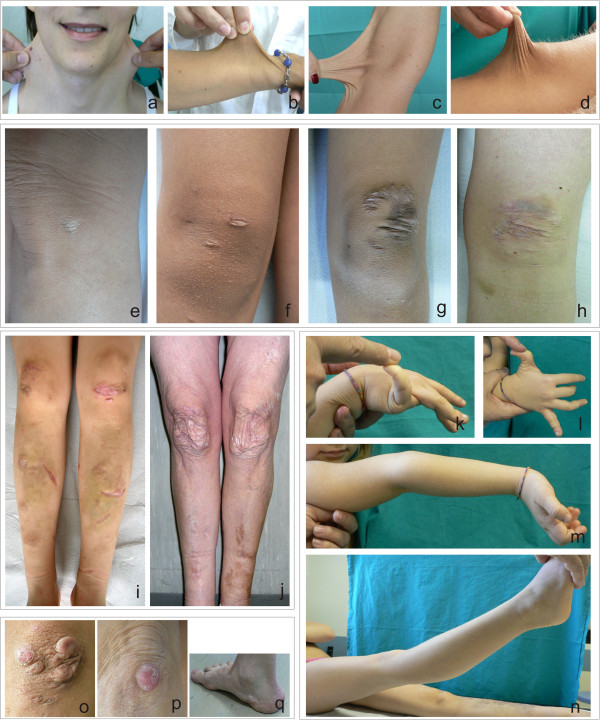
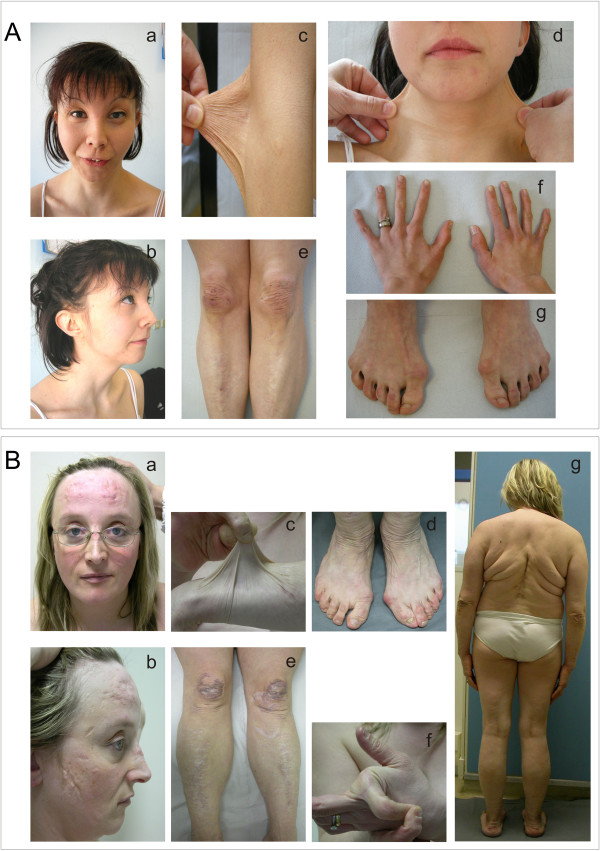
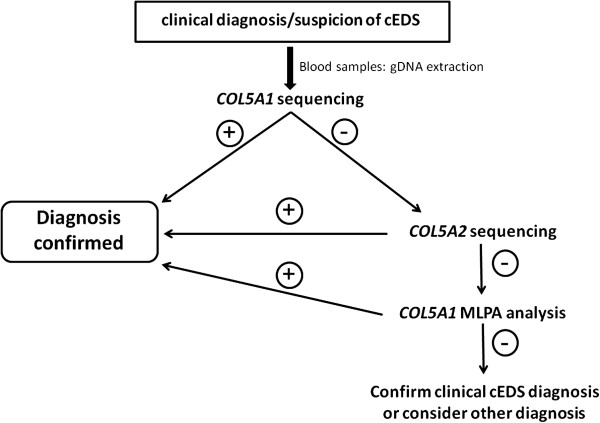
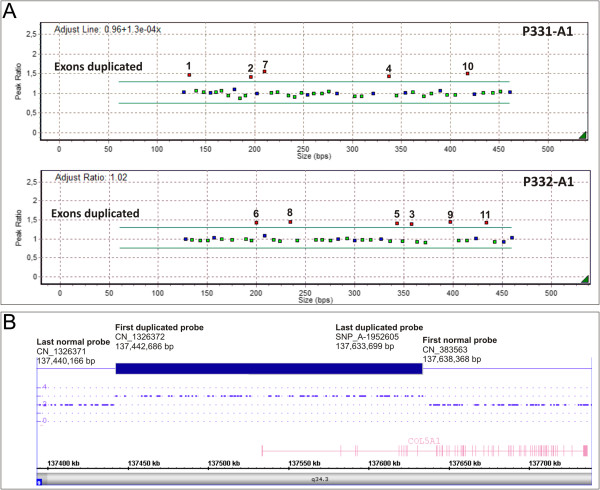
Background: Classic Ehlers–Danlos syndrome (cEDS) is a rare autosomal dominant connective tissue disorder that is primarily characterized by skin hyperextensibility, abnormal wound healing/atrophic scars, and joint hypermobility. A recent study demonstrated that more than 90% of patients who satisfy all of these major criteria harbor a type V collagen (COLLV) defect. Methods: This cohort included 40 patients with cEDS who were clinically diagnosed according to the Villefranche nosology. The flowchart that was adopted for mutation detection consisted of sequencing the COL5A1 gene and, if no mutation was detected, COL5A2 analysis. In the negative patients the presence of large genomic rearrangements in COL5A1 was investigated using MLPA, and positive results were confirmed via SNP-array analysis. Results: We report the clinical and molecular characterization of 40 patients from 28 families, consisting of 14 pediatric patients and 26 adults. A family history of cEDS was present in 9 patients. The majority of the patients fulfilled all the major diagnostic criteria for cEDS; atrophic scars were absent in 2 females, skin hyperextensibility was not detected in a male and joint hypermobility was negative in 8 patients (20% of the entire cohort). Wide inter- and intra-familial phenotypic heterogeneity was observed. We identified causal mutations with a detection rate of approximately 93%. In 25/28 probands, COL5A1 or COL5A2 mutations were detected. Twenty-one mutations were in the COL5A1 gene, 18 of which were novel (2 recurrent). Of these, 16 mutations led to nonsense-mediated mRNA decay (NMD) and to COLLV haploinsufficiency and 5 mutations were structural. Two novel COL5A2 splice mutations were detected in patients with the most severe phenotypes. The known p. (Arg312Cys) mutation in the COL1A1 gene was identified in one patient with vascular-like cEDS. Conclusions: Our findings highlight that the three major criteria for cEDS are useful and sufficient for cEDS clinical diagnosis in the large majority of the patients. The borderline patients for whom these criteria fail can be diagnosed when minor signs of connective tissue diseases and family history are present and when genetic testing reveals a defect in COLLV. Our data also confirm that COL5A1 and COL5A2 are the major, if not the only, genes involved in cEDS.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“Morethan 90% of cEDS patients harbor a heterozygous mutation in one of the genes encoding type V collagen ( COL5A1 and COL5A2 ) [Symoens et al., 2012; Ritelli et al., 2013; Zoppi et al., 2015]”
From The 2017 international classification of the Ehlers–Danlos syndromes (2017) · cites this paper as Ritelli et al., 2013
The cited source says what the claim attributes to it.
In the citing paper
“The p.(Arg312Cys) mutation has subsequently been identified in two other individuals with EDS and complications of vascular fragility [Ritelli et al., 2013;Gaines et al., 2015].”
From The Ehlers–Danlos syndromes, rare types (2017) · cites this paper as Ritelli et al., 2013
The cited source says what the claim attributes to it.
In the citing paper
“Patients with the p.(Arg312Cys) mutation are particularly at risk for vascular events, highlighting the benefit of molecular confirmation in classical EDS patients for management purposes.”
From The Ehlers–Danlos syndromes, rare types (2017) · cites this paper as Ritelli et al., 2013
In the citing paper
“cEDS and vEDS are caused respectively by defects in collagen V and collagen III, whereas the molecular basis of hEDS is unknown.”
From Cellular and Molecular Mechanisms in the Pathogenesis of Classical, Vascular, and Hypermobile Ehlers‒Danlos Syndromes (2019) · cites this paper as [41]
In the citing paper
“Collagen V haploinsufficiency is the most common molecular defect caused by COL5A1 null alleles, whereas rare COL5A1 variants and the majority of COL5A2 mutations reported so far affect collagen V structural integrity by exerting a dominant negative effect [40,41].”
From Cellular and Molecular Mechanisms in the Pathogenesis of Classical, Vascular, and Hypermobile Ehlers‒Danlos Syndromes (2019) · cites this paper as [41]
In the citing paper
“Collagen V haploinsufficiency is the most common molecular defect caused by COL5A1 null alleles, whereas rare COL5A1 variants and the majority of COL5A2 mutations reported so far affect collagen V structural integrity by exerting a dominant negative effect [40,41].”
From Cellular and Molecular Mechanisms in the Pathogenesis of Classical, Vascular, and Hypermobile Ehlers‒Danlos Syndromes (2019) · cites this paper as [41]
In the citing paper
“Most patients harbor point mutations or chromosomal rearrangements in COL5A1 or COL5A2 genes encoding the collagen V [40,41].”
From Cellular and Molecular Mechanisms in the Pathogenesis of Classical, Vascular, and Hypermobile Ehlers‒Danlos Syndromes (2019) · cites this paper as [41]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Venselaar, H., Beek, T. A. H. T., Kuipers, R. K. P., Hekkelman, M. L., & Vriend, G. (2010) · BMC Bioinformatics · doi:10.1186/1471-2105-11-548 · Cited by 1,163
Type V collagen: heterotypic type I/V collagen interactions in the regulation of fibril assembly
Birk, D. E. (2001) · Micron · doi:10.1016/s0968-4328(00)00043-3 · Cited by 438 · cited 2×
“nents that are involved in the maintenance of ECM architecture [12-16].”
BackgroundA recessive form of the Ehlers-Danlos syndrome caused by tenascin-X deficiency
Schalkwijk, J., Zweers, M. C., Steijlen, P. M., Dean, W. B., Taylor, G., van Vlijmen, I. M., van Haren, B., Miller, W. L., & Bristow, J. (2001) · New England Journal of Medicine · doi:10.1056/nejmoa002939 · Cited by 391 · cited 1×
“Moreover, the associated phenotypes exhibit certain differences from those of COLLV-associated cEDS [11,36,37].”
The Ehlers–Danlos syndrome, a disorder with many faces
De Paepe, A., & Malfait, F. (2012) · Clinical Genetics · doi:10.1111/j.1399-0004.2012.01858.x · Cited by 379 · cited 4×
“Aortic ring dilatation, large-size arterial aneurysms and dissections, intracranial aneurysms, and arterio-venous fistulas have rarely been observed [3-9].”
BackgroundInfluentialSteinmann, B., Royce, P. M., & Superti‐Furga, A. (2002) · Connective Tissue and Its Heritable Disorders · doi:10.1002/0471221929.ch9 · Cited by 358
Clinical and genetic aspects of Ehlers-Danlos syndrome, classic type
Malfait, F., WENSTRUP, R. J., & Paepe, A. D. (2010) · Genetics in Medicine · doi:10.1097/gim.0b013e3181eed412 · Cited by 303
The Human Collagen Mutation Database 1998
Dalgleish, R. (1998) · Nucleic Acids Research · doi:10.1093/nar/26.1.253 · Cited by 219
Classical Ehlers-Danlos Syndrome Caused by a Mutation in Type I Collagen
Nuytinck, L., Freund, M., Lagae, L., Pierard, G. E., Hermanns-Le, T., & De Paepe, A. (2000) · The American Journal of Human Genetics · doi:10.1086/302859 · Cited by 169 · cited 2×
“Moreover, the associated phenotypes exhibit certain differences from those of COLLV-associated cEDS [11,36,37].”
BackgroundMalfait, F., Symoens, S., De Backer, J., Hermanns-Lê, T., Sakalihasan, N., Lapière, C. M., Coucke, P., & De Paepe, A. (2007) · Human Mutation · doi:10.1002/humu.20455 · Cited by 167 · cited 2×
“young adulthood [36,38].”
Mutations in the COL5A1 gene are causal in the Ehlers-Danlos syndromes I and II.
Paepe, A. D., Nuytinck, L., Haußer, I., Anton‐Lamprecht, I., & Naeyaert, J. M. (1997) · Europe PMC (PubMed Central) · Cited by 165
Research works printed but not linked: 1
- Levy HP: Ehlers-Danlos syndrome, hypermobility type. GeneReviews™ [Internet]. Seattle (WA). Edited by: Pagon RA, Bird TD, Dolan CR, Stephens K, Adam MP. 1993, Seattle: University of Washington, updated 2012 Sep 13
Cited by
Sorted by citations
The 2017 international classification of the Ehlers–Danlos syndromes
Malfait, F., Francomano, C., Byers, P., Belmont, J., Berglund, B., Black, J., Bloom, L., Bowen, J. M., Brady, A. F., Burrows, N. P., et al. (2017) · American Journal of Medical Genetics Part C: Seminars in Medical Genetics · doi:10.1002/ajmg.c.31552 · Cited by 1,936
“• Molecular basis More than 90% of cEDS patients harbor a heterozygous mutation in one of the genes encoding type V collagen (COL5A1 and COL5A2) [Symoens et al., 2012; Ritelli et al., 2013; Zoppi et al., 2015] (see also “Ehlers–Danlos Syndrome, Classical Type,” by Bowen et al., this issue).”
BackgroundSelf-citationLattouf, R., Younes, R., Lutomski, D., Naaman, N., Godeau, G., Senni, K., & Changotade, S. (2014) · Journal of Histochemistry & Cytochemistry · doi:10.1369/0022155414545787 · Cited by 524
“COL5A1/COL5A2 mutations (Symoens et al. 2012; Ritelli et al. 2013).”
Malfait, F., Castori, M., Francomano, C. A., Giunta, C., Kosho, T., & Byers, P. H. (2020) · Nature Reviews Disease Primers · doi:10.1038/s41572-020-0194-9 · Cited by 342
The Ehlers–Danlos syndromes, rare types
Brady, A. F., Demirdas, S., Fournel‐Gigleux, S., Ghali, N., Giunta, C., Kapferer‐Seebacher, I., Kosho, T., Mendoza‐Londono, R., Pope, M. F., Rohrbach, M., et al. (2017) · American Journal of Medical Genetics Part C Seminars in Medical Genetics · doi:10.1002/ajmg.c.31550 · Cited by 258
“[Nuytinck et al., 2000; Malfait et al., 2007; Ritelli et al., 2013; Gaines et al., 2015].”
BackgroundEhlers-Danlos syndrome, classical type
Bowen, J. M., Sobey, G. J., Burrows, N. P., Colombi, M., Lavallee, M. E., Malfait, F., & Francomano, C. A. (2017) · American Journal of Medical Genetics Part C: Seminars in Medical Genetics · doi:10.1002/ajmg.c.31548 · Cited by 184
“Advances in molecular testing have allowed more type V collagen abnormalities to be identified, and recent reports indicate that the majority of patients with classical EDS do have a mutation in either COL5A1 or COL5A2 [Symoens et al., 2012; Ritelli et al., 2013].”
BackgroundInfluentialSelf-citation
Metadata
- Authors
- Ritelli, M., Dordoni, C., Venturini, M., Chiarelli, N., Quinzani, S., Traversa, M., Zoppi, N., Vascellaro, A., Wischmeijer, A., Manfredini, E., Garavelli, L., Calzavara-Pinton, P., & Colombi, M.
- Affiliations
- University of Brescia (IT)
- Azienda Socio Sanitaria Territoriale degli Spedali Civili di Brescia (IT)
- IRCCS Azienda Ospedliero-Universitaria di Bologna Policlinico di Sant'Orsola (IT)
- Azienda Socio Sanitaria Territoriale Grande Ospedale Metropolitano Niguarda (IT)
- Santa Maria Nuova Hospital (IT)
- Corresponding author
- Colombi, Marina
- ORCID
- Venue
- Orphanet Journal of Rare Diseases
- Published
- 2013-04-12
- Volume
- 8
- Issue
- 1
- Pages
- 58-58
- ISSN
- 1750-1172 (ISSN-L)
- PMC
- PMC3653713
- OpenAlex
- W2065067440
- Cited by
- 134
- References
- 40
- Citations by year
- 2026: 8 · 2025: 7 · 2024: 6 · 2023: 3 · 2022: 8 · 2021: 15 · 2020: 14 · 2019: 15 · 2018: 14 · 2017: 12 · 2016: 12 · 2015: 9 · 2014: 9 · 2013: 2
- FWCI
- 7.08
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- cc-by
- Locations
- Orphanet Journal of Rare Diseases (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Institutional Research Information System (Università degli Studi di Brescia) (submittedVersion, other-oa)
- Funders
- Regione Lombardia
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Connective tissue disorders research
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genomics and Rare Diseases
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Dermatological and Skeletal Disorders
- Keywords
- type V collagen, novel mutations, nonsense-mediated mRNA decay, splicing mutation, phenotypic heterogeneity, connective tissue diseases, vascular Ehlers-Danlos syndrome
- MeSH terms
- Adolescent, Adult, Child, Collagen, Ehlers-Danlos Syndrome, Family, Female, Humans, Male, Mutation, Collagen Type V, Haploinsufficiency, Nonsense Mediated mRNA Decay
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
