

The DOI matches its Crossref record; 31 of the 31 references it cites were found in a registry or an index.
Simultaneous Analysis of All SNPs in Genome-Wide and Re-Sequencing Association Studies
Open accessAbstract
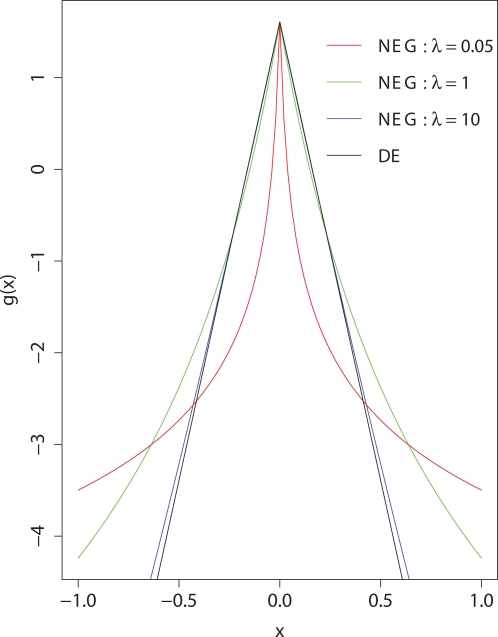
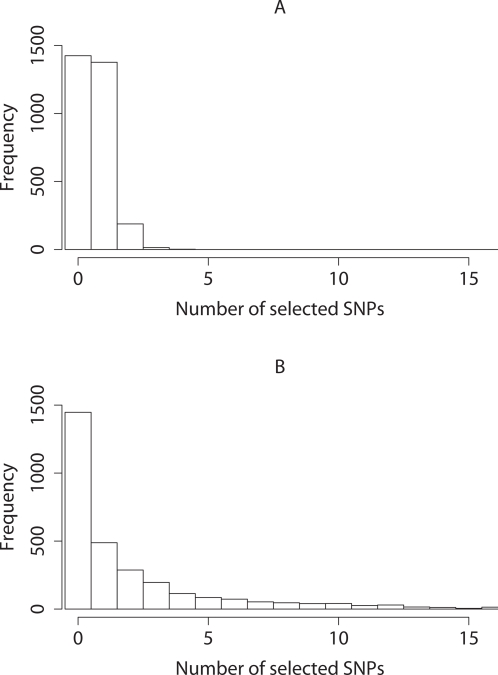
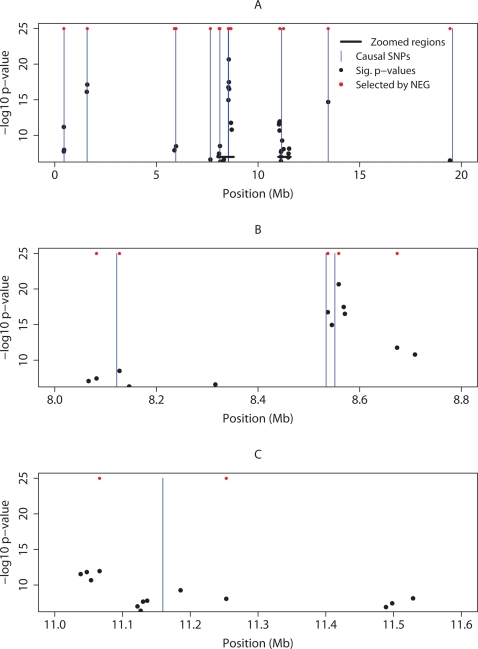
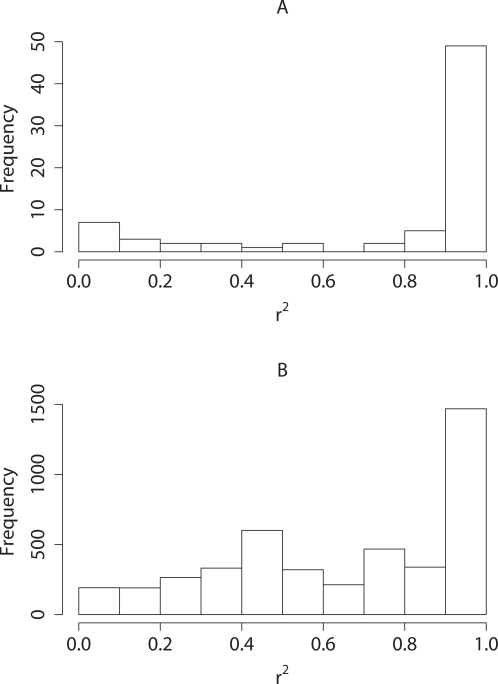
Testing one SNP at a time does not fully realise the potential of genome-wide association studies to identify multiple causal variants, which is a plausible scenario for many complex diseases. We show that simultaneous analysis of the entire set of SNPs from a genome-wide study to identify the subset that best predicts disease outcome is now feasible, thanks to developments in stochastic search methods. We used a Bayesian-inspired penalised maximum likelihood approach in which every SNP can be considered for additive, dominant, and recessive contributions to disease risk. Posterior mode estimates were obtained for regression coefficients that were each assigned a prior with a sharp mode at zero. A non-zero coefficient estimate was interpreted as corresponding to a significant SNP. We investigated two prior distributions and show that the normal-exponential-gamma prior leads to improved SNP selection in comparison with single-SNP tests. We also derived an explicit approximation for type-I error that avoids the need to use permutation procedures. As well as genome-wide analyses, our method is well-suited to fine mapping with very dense SNP sets obtained from re-sequencing and/or imputation. It can accommodate quantitative as well as case-control phenotypes, covariate adjustment, and can be extended to search for interactions. Here, we demonstrate the power and empirical type-I error of our approach using simulated case-control data sets of up to 500 K SNPs, a real genome-wide data set of 300 K SNPs, and a sequence-based dataset, each of which can be analysed in a few hours on a desktop workstation.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
In the citing paper
“Simulation studies and permutation testing that consider the entire set of tests performed (e.g., all windows and groups of variants across all genomic regions considered) to a get a global false positive rate are the most appropriate given their flexibility and sound theoretical bases, but will likely be very computationally intensive.75”
From Statistical analysis strategies for association studies involving rare variants (2010) · cites this paper as [75]
In the citing paper
“Newer regression techniques that make use of regularization and shrinkage parameters to control for collinearity and overfitting can be used to overcome these problems.”
From Statistical analysis strategies for association studies involving rare variants (2010) · cites this paper as [75]
In the citing paper
“Two such techniques, ridge regression74 and the LASSO75,76 have been considered in genetic association analysis contexts, and other methods have also been proposed as well.77–80, 81”
From Statistical analysis strategies for association studies involving rare variants (2010) · cites this paper as [75]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Regression shrinkage and selection via the lasso
Tibshirani, R. (1996) · Journal of the Royal Statistical Society Series B: Statistical Methodology · doi:10.1111/j.2517-6161.1996.tb02080.x · Cited by 51,482
Genome-wide association study of 14,000 cases of seven common diseases and 3,000 shared controls
The Wellcome Trust Case Control Consortium, Management Committee, Burton, P. R., Clayton, D. G., Cardon, L. R., Craddock, N., Deloukas, P., Duncanson, A., Kwiatkowski, D. P., McCarthy, M. I., et al. (2007) · Nature · doi:10.1038/nature05911 · Cited by 9,442
Prediction of Total Genetic Value Using Genome-Wide Dense Marker Maps
Meuwissen, T. H. E., Hayes, B. J., & Goddard, M. E. (2001) · Genetics · doi:10.1093/genetics/157.4.1819 · Cited by 7,917
Nonlinear programming: theory and algorithms
(1993) · Computers & Operations Research · doi:10.1016/0305-0548(93)90042-h · Cited by 5,803
Population Structure and Eigenanalysis
Patterson, N., Price, A. L., & Reich, D. (2006) · PLoS Genetics · doi:10.1371/journal.pgen.0020190 · Cited by 5,280
A genome-wide association study identifies novel risk loci for type 2 diabetes
Sladek, R., Rocheleau, G., Rung, J., Dina, C., Shen, L., Serre, D., Boutin, P., Vincent, D., Belisle, A., Hadjadj, S., et al. (2007) · Nature · doi:10.1038/nature05616 · Cited by 2,993
Self-citationVariable Selection via Gibbs Sampling
George, E. I., & McCulloch, R. E. (1993) · Journal of the American Statistical Association · doi:10.1080/01621459.1993.10476353 · Cited by 2,828
The Wellcome Trust Case Control Consortium, U.K.
Mossman, K. (2008) · Scientific American · doi:10.1038/scientificamerican0108-42a · Cited by 2,714
Bayesian Variable Selection in Linear Regression
Mitchell, T. J., & Beauchamp, J. J. (1988) · Journal of the American Statistical Association · doi:10.1080/01621459.1988.10478694 · Cited by 1,427
(1997) · Cited by 1,193
Research works printed but not linked: 1
- Y Li, 2006, Mach 1.0: Rapid haplotype reconstruction and missing geno-type inference., Am J Hum Genet, S79, 2290
Cited by
Sorted by citations
Items 101–200 of 321
Lin, W., Camp, N. J., Ghoussaini, M., Beesley, J., Michailidou, K., Hopper, J. L., Apicella, C., Southey, M. C., Stone, J., Schmidt, M. K., et al. (2014) · Human Molecular Genetics · doi:10.1093/hmg/ddu431 · Cited by 45
A review of SNP heritability estimation methods
Tang, M., Wang, T., & Zhang, X. (2022) · Briefings in Bioinformatics · doi:10.1093/bib/bbac067 · Cited by 46
Pośpiech, E., Kukla‐Bartoszek, M., Karłowska-Pik, J., Zieliński, P., Woźniak, A., Boroń, M., Dąbrowski, M., Zubańska, M., Jarosz, A., Grzybowski, T., et al. (2020) · BMC Genomics · doi:10.1186/s12864-020-06926-y · Cited by 42
Yang, C., Wan, X., Yang, Q., Xue, H., & Yu, W. (2010) · BMC Bioinformatics · doi:10.1186/1471-2105-11-s1-s18 · Cited by 42
A comparison of methods for whole-genome QTL mapping using dense markers in four livestock species
Legarra, A., Croiseau, P., Sanchez, M., Teyssèdre, S., Sallé, G., Allais, S., Fritz, S., Moreno, C., Ricard, A., & Elsen, J.-M. (2015) · Genetics Selection Evolution · doi:10.1186/s12711-015-0087-7 · Cited by 39
Applying compressed sensing to genome-wide association studies
Vattikuti, S., Lee, J. J., Chang, C. C., Hsu, S., & Chow, C. C. (2014) · GigaScience · doi:10.1186/2047-217x-3-10 · Cited by 38
Fast and flexible joint fine-mapping of multiple traits via the Sum of Single Effects model
Zou, Y., Carbonetto, P. S., Xie, D., Wang, G., & Stephens, M. (2026) · Nature Genetics · doi:10.1038/s41588-025-02486-7 · Cited by 44
Barrett, J. H., Taylor, J., Bright, C. J., Harland, M., Dunning, A. M., Akslen, L. A., Andresen, P. A., Avril, M., Azizi, E., Bianchi‐Scarrà, G., et al. (2014) · International Journal of Cancer · doi:10.1002/ijc.29099 · Cited by 38
PUMA: A Unified Framework for Penalized Multiple Regression Analysis of GWAS Data
Hoffman, G. E., Logsdon, B. A., & Mezey, J. G. (2013) · PLoS Computational Biology · doi:10.1371/journal.pcbi.1003101 · Cited by 40
Denis, M., Favreau, B., Ueno, S., Camus‐Kulandaivelu, L., Chaix, G., Gion, J., Nourrisier-Mountou, S., Polidori, J., & Bouvet, J. M. (2013) · Tree Genetics & Genomes · doi:10.1007/s11295-013-0606-z · Cited by 39
Panning for Gold: Model-X Knockoffs for High-dimensional Controlled Variable SelectionPreprint
Candès, E. J., Fan, Y., Janson, L., & Lv, J. (2016) · arXiv (Cornell University) · doi:10.48550/arxiv.1610.02351 · Cited by 39
Du, Q., Tian, J., Yang, X., Pan, W., Xu, B., Li, B., Ingvarsson, P. K., & Zhang, D. (2014) · DNA Research · doi:10.1093/dnares/dsu040 · Cited by 40
Genome-Wide Association Study Statistical Models: A Review
Yoosefzadeh-Najafabadi, M., Eskandari, M., Belzile, F. J., & Torkamaneh, D. (2022) · Methods in molecular biology · doi:10.1007/978-1-0716-2237-7_4 · Cited by 40
SparSNP: Fast and memory-efficient analysis of all SNPs for phenotype prediction
Abraham, G., Kowalczyk, A., Zobel, J., & Inouye, M. (2012) · BMC Bioinformatics · doi:10.1186/1471-2105-13-88 · Cited by 37
Fang, M., Fu, W., Jiang, D. L., Zhang, Q., Sun, D., Ding, X., & Liu, J. (2014) · PLoS ONE · doi:10.1371/journal.pone.0099544 · Cited by 36
Statistical analysis of genetic interactions
Yi, N. (2010) · Genetics Research · doi:10.1017/s0016672310000595 · Cited by 36
Lee, S. H., Nyholt, D. R., Macgregor, S., Henders, A. K., Zondervan, K. T., Montgomery, G. W., & Visscher, P. M. (2010) · Genetic Epidemiology · doi:10.1002/gepi.20541 · Cited by 33
Robustness of Bayesian Multilocus Association Models to Cryptic Relatedness
Kärkkäinen, H., & Sillanpää, M. J. (2012) · Annals of Human Genetics · doi:10.1111/j.1469-1809.2012.00729.x · Cited by 34
Chenevix‐Trench, G., Carvajal‐Carmona, L. G., Chenevix‐Trench, G., RENDOCAS, (AOCS), A. O. C. S., The GENICA Network, O’Mara, T. A., Painter, J. N., Lose, F. A., Dennis, J., et al. (2014) · Human Genetics · doi:10.1007/s00439-014-1515-4 · Cited by 34
Probabilistic Modeling of Imaging, Genetics and Diagnosis
Batmanghelich, K. N., Dalca, A. V., Quon, G., Sabuncu, M. R., & Golland, P. (2016) · IEEE Transactions on Medical Imaging · doi:10.1109/tmi.2016.2527784 · Cited by 36
Bag of Naïve Bayes: biomarker selection and classification from genome-wide SNP data
Sambo, F., Trifoglio, E., Camillo, B. D., TOFFOLO, G. M., & Cobelli, C. (2012) · BMC Bioinformatics · doi:10.1186/1471-2105-13-s14-s2 · Cited by 34
Analysis and Optimization of Bulk DNA Sampling with Binary Scoring for Germplasm Characterization
Reyes‐Valdés, M. H., Santacruz‐Varela, A., Martínez, O., Simpson, J. K., Hayano‐Kanashiro, C., & Cortés-Romero, C. (2013) · PLoS ONE · doi:10.1371/journal.pone.0079936 · Cited by 34
Ultrahigh-dimensional variable selection method for whole-genome gene-gene interaction analysis
Ueki, M., & Tamiya, G. (2012) · BMC Bioinformatics · doi:10.1186/1471-2105-13-72 · Cited by 34
A semi-automatic method to guide the choice of ridge parameter in ridge regressionPreprint
Cule, E., & Iorio, M. D. (2012) · arXiv (Cornell University) · doi:10.48550/arxiv.1205.0686 · Cited by 33
Detecting Genetic Interactions in Pathway‐Based Genome‐Wide Association Studies
Huang, A., Martin, E. R., Vance, J. M., & Cai, X. (2014) · Genetic Epidemiology · doi:10.1002/gepi.21803 · Cited by 32
Denis, M., Enquobahrie, D. A., Tadesse, M. G., Gelaye, B., Sánchez, S. E., Salazar, M., Ananth, C. V., & Williams, M. A. (2014) · PLoS ONE · doi:10.1371/journal.pone.0116346 · Cited by 32
Yi, N., Kaklamani, V., & Pasche, B. (2010) · Annals of Human Genetics · doi:10.1111/j.1469-1809.2010.00605.x · Cited by 31
Genomic prediction of celiac disease targeting HLA-positive individuals
Abraham, G., Rohmer, A., Tye–Din, J. A., & Inouye, M. (2015) · Genome Medicine · doi:10.1186/s13073-015-0196-5 · Cited by 30
Confounded by sequencing depth in association studies of rare alleles
Garner, C. P. (2011) · Genetic Epidemiology · doi:10.1002/gepi.20574 · Cited by 31
Association of the IL-10 Gene Family Locus on Chromosome 1 with Juvenile Idiopathic Arthritis (JIA)
Omoyinmi, E., Forabosco, P., Hamaoui, R., Bryant, A., Hinks, A., Ursu, S., (CAPS), C. A. P. S., BSPAR study group, (CHARMS), C. A. R. T. M. S., Wedderburn, L. R., et al. (2012) · PLoS ONE · doi:10.1371/journal.pone.0047673 · Cited by 31
Advances in the Pharmacogenomics of Adverse Drug Reactions
Collins, S. L., Carr, D. F., & Pirmohamed, M. (2015) · Drug Safety · doi:10.1007/s40264-015-0367-8 · Cited by 31
RWEN: response-weighted elastic net for prediction of chemosensitivity of cancer cell lines
Basu, A. B., Mitra, R., Liu, H., Schreiber, S. L., & Clemons, P. A. (2018) · Bioinformatics · doi:10.1093/bioinformatics/bty199 · Cited by 32
Chinn, L. W., Tang, M., Kessing, B. D., Lautenberger, J. A., Troyer, J. L., Malasky, M., MCINTOSH, C. E., Kirk, G. D., Wolinsky, S. M., Buchbinder, S., et al. (2010) · The Journal of Infectious Diseases · doi:10.1086/657322 · Cited by 28
Detection boundary and Higher Criticism approach for rare and weak genetic effects
Wu, Z., Sun, Y., He, S., Cho, J. H., Zhao, H., & Jin, J. (2014) · The Annals of Applied Statistics · doi:10.1214/14-aoas724 · Cited by 28
Takahashi, Y., Ueki, M., Tamiya, G., Ogishima, S., Kinoshita, K., Hozawa, A., Minegishi, N., Nagami, F., Fukumoto, K., Otsuka, K., et al. (2020) · Translational Psychiatry · doi:10.1038/s41398-020-00957-5 · Cited by 29
Peng, B., Zhu, D., Ander, B. P., Zhang, X., Xue, F., Sharp, F. R., & Yang, X. (2013) · PLoS ONE · doi:10.1371/journal.pone.0067672 · Cited by 28
Whole-Genome Quantitative Trait Locus Mapping Reveals Major Role of Epistasis on Yield of Rice
Huang, A., Xu, S., & Cai, X. (2014) · PLoS ONE · doi:10.1371/journal.pone.0087330 · Cited by 26
SNP‐based heritability and selection analyses: Improved models and new results
Speed, D., Kaphle, A., & Balding, D. J. (2022) · BioEssays · doi:10.1002/bies.202100170 · Cited by 26
Self-citationReprioritizing Genetic Associations in Hit Regions Using LASSO‐Based Resample Model Averaging
Valdar, W. S. J., Sabourin, J., Nobel, A. B., & Holmes, C. C. (2012) · Genetic Epidemiology · doi:10.1002/gepi.21639 · Cited by 26
Empirical Bayesian LASSO-logistic regression for multiple binary trait locus mapping
Huang, A., Xu, S., & Cai, X. (2013) · BMC Genetics · doi:10.1186/1471-2156-14-5 · Cited by 25
Wang, Y., Chen, J., Guan, Z., Zhang, X., Zhang, Y., Ma, L., Yao, Y., Peng, H., Zhang, Q., Zhang, B., et al. (2019) · Molecular Genetics and Genomics · doi:10.1007/s00438-019-01586-4 · Cited by 26
Alisoltani, A., Fallahi, H., Ebrahimi, M. M., Ebrahimi, M., & Ebrahimie, E. (2014) · PLoS ONE · doi:10.1371/journal.pone.0096320 · Cited by 23
Bayesian Sparsity-Path-Analysis of Genetic Association Signal using Generalized t Priors
Lee, A., Caron, F., Arnaud Doucet, B., & Holmes, C. C. (2012) · Statistical Applications in Genetics and Molecular Biology · doi:10.2202/1544-6115.1712 · Cited by 23
Hierarchical inference for genome-wide association studies: a view on methodology with software
Renaux, C., Buzdugan, L., Kalisch, M., & Bühlmann, P. (2020) · Computational Statistics · doi:10.1007/s00180-019-00939-2 · Cited by 21
Toosi, A., Fernando, R. L., & Dekkers, J. C. M. (2018) · Genetics Selection Evolution · doi:10.1186/s12711-018-0402-1 · Cited by 23
netgwas: An R Package for Network-Based Genome Wide Association Studies
Behrouzi, P., Arends, D., & Wit, E. C. (2023) · The R Journal · doi:10.32614/rj-2023-011 · Cited by 23
Genetics of Complex Airway Disease
Cookson, W., & Moffatt, M. F. (2011) · Proceedings of the American Thoracic Society · doi:10.1513/pats.201101-003ms · Cited by 21
Bioinformatics Methods to Select Prognostic Biomarker Genes from Large Scale Datasets: A Review
Jardillier, R., Chatelain, F., & Guyon, L. (2018) · Biotechnology Journal · doi:10.1002/biot.201800103 · Cited by 21
Lu, Z., Hongtu Zhu, B., Knickmeyer, R. C., Sullivan, P. F., Williams, S., Zou, F., & Initiative, A. D. N. (2015) · Genetic Epidemiology · doi:10.1002/gepi.21932 · Cited by 22
Darabi, H., Beesley, J., Droit, A., Kar, S., Nord, S., Marjaneh, M. M., Soucy, P., Michailidou, K., Ghoussaini, M., Wahl, H. F., et al. (2016) · Scientific Reports · doi:10.1038/srep32512 · Cited by 22
Using biological knowledge to discover higher order interactions in genetic association studies
Chen, G. K., & Thomas, D. C. (2010) · Genetic Epidemiology · doi:10.1002/gepi.20542 · Cited by 22
A hybrid bayesian approach for genome-wide association studies on related individuals
Yazdani, A., & Dunson, D. B. (2015) · Bioinformatics · doi:10.1093/bioinformatics/btv496 · Cited by 20
Ehsani, A., Sørensen, P., Pomp, D., Allan, M. F., & Janss, L. (2012) · BMC Genomics · doi:10.1186/1471-2164-13-456 · Cited by 19
Genome wide association study identifies a novel putative mammographic density locus at 1q12‐q21
Fernández‐Navarro, P., González‐Neira, A., Pita, G., Díaz‐Uriarte, R., Moreno, L. T., Ederra, M., Pedraz‐Pingarrón, C., Sánchez‐Contador, C., Vázquez-Carrete, J. A., Moreo, P., et al. (2014) · International Journal of Cancer · doi:10.1002/ijc.29299 · Cited by 21
Dolejsi, E., Bodenstorfer, B., & Frommlet, F. (2014) · PLoS ONE · doi:10.1371/journal.pone.0103322 · Cited by 20
Bertrand, J., & Balding, D. J. (2013) · Pharmacogenetics and Genomics · doi:10.1097/fpc.0b013e32835dd22c · Cited by 20
Self-citationLogsdon, B. A., Carty, C. L., Reiner, A. P., Dai, J. Y., & Kooperberg, C. L. (2012) · Bioinformatics · doi:10.1093/bioinformatics/bts261 · Cited by 19
Genetic Prediction of Quantitative Lipid Traits: Comparing Shrinkage Models to Gene Scores
Warren, H. R., Casas, J. P., Hingorani, A. D., Dudbridge, F., & Whittaker, J. C. (2013) · Genetic Epidemiology · doi:10.1002/gepi.21777 · Cited by 20
Self-citationA Longitudinal HbA1c Model Elucidates Genes Linked to Disease Progression on Metformin
Goswami, S., Yee, S. W., Xu, F., Sridhar, S., Mosley, J. D., Takahashi, A., Kubo, M., Maeda, S., Davis, R., Roden, D. M., et al. (2016) · Clinical Pharmacology & Therapeutics · doi:10.1002/cpt.428 · Cited by 20
Prediction of treatment response in rheumatoid arthritis patients using genome‐wide SNP data
Cherlin, S., Plant, D., Taylor, J., Colombo, M., Spiliopoulou, A., Tzanis, E., Morgan, A. W., Barnes, M. R., McKeigue, P., Barrett, J. H., et al. (2018) · Genetic Epidemiology · doi:10.1002/gepi.22159 · Cited by 20
Using Penalised Logistic Regression to Fine Map HLA Variants for Rheumatoid Arthritis
Vignal, C., Bansal, A. T., & Balding, D. J. (2011) · Annals of Human Genetics · doi:10.1111/j.1469-1809.2011.00670.x · Cited by 20
Self-citationA comparison of whole genome sequencing with exome sequencing for family-based association studies
Lacey, S., Chung, J., & Lin, H. (2014) · BMC Proceedings · doi:10.1186/1753-6561-8-s1-s38 · Cited by 19
Li, P., Wang, Y., Cheng, J., Chen, J., & Ha, M.-W. (2015) · Tumor Biology · doi:10.1007/s13277-015-3672-z · Cited by 18
Dai, X., Fu, G., Zhao, S., & Zeng, Y. (2021) · Genes · doi:10.3390/genes12050736 · Cited by 18
Bayesian variable selection for hierarchical gene–environment and gene–gene interactions
Liu, C., Ma, J., & Amos, C. I. (2014) · Human Genetics · doi:10.1007/s00439-014-1478-5 · Cited by 18
Hoffman, G. E. (2013) · PLoS ONE · doi:10.1371/annotation/31412345-fc86-4d67-b37c-93d42f5f0a59 · Cited by 18
Adelson, R. P., Renton, A. E., Li, W., Barzilai, N., Atzmon, G., Goate, A., Davies, P., & Freudenberg‐Hua, Y. (2019) · Scientific Reports · doi:10.1038/s41598-019-52614-7 · Cited by 17
Li, Y., & Kim, J.-J. (2015) · Asian-Australasian Journal of Animal Sciences · doi:10.5713/ajas.15.0077 · Cited by 17
Functional annotation signatures of disease susceptibility loci improve SNP association analysis
Iversen, E. S., Lipton, G. B., Clyde, M. A., & Monteiro, Á. N. A. (2014) · BMC Genomics · doi:10.1186/1471-2164-15-398 · Cited by 17
Hettiarachchi, G. K., & Komar, A. A. (2022) · Springer eBooks · doi:10.1007/978-3-031-05616-1_4 · Cited by 17
A Poisson reduced-rank regression model for association mapping in sequencing data
Fitzgerald, T., Jones, A., & Engelhardt, B. E. (2022) · BMC Bioinformatics · doi:10.1186/s12859-022-05054-6 · Cited by 18
Bansal, V., Libiger, O., Torkamani, A., & Schork, N. J. (2010) · WORLD SCIENTIFIC eBooks · doi:10.1142/9789814335058_0009 · Cited by 16
Multilocus association mapping using generalized ridge logistic regression
Liu, Z., Shen, Y., & Ott, J. (2011) · BMC Bioinformatics · doi:10.1186/1471-2105-12-384 · Cited by 16
Peltola, T., Marttinen, P., & Vehtari, A. (2012) · PLoS ONE · doi:10.1371/journal.pone.0049445 · Cited by 16
Khondoker, M. R., Newhouse, S., Westman, E., Muehlboeck, J., Mecocci, P., Vellas, B. J., Tsolaki, M., Kłoszewska, I., Soininen, H. S., Lovestone, S., et al. (2015) · Journal of Alzheimer s Disease · doi:10.3233/jad-142214 · Cited by 16
A novel algorithm for simultaneous SNP selection in high-dimensional genome-wide association studies
Zuber, V., Silva, A. P. D., & Strimmer, K. (2012) · BMC Bioinformatics · doi:10.1186/1471-2105-13-284 · Cited by 16
EBglmnet: a comprehensive R package for sparse generalized linear regression models
Huang, A., & Liu, D. (2016) · Bioinformatics · doi:10.1093/bioinformatics/btw143 · Cited by 16
Structured Genome-Wide Association Studies with Bayesian Hierarchical Variable Selection
Zhao, Y., Hongtu Zhu, B., Lu, Z., Knickmeyer, R. C., & Zou, F. (2019) · Genetics · doi:10.1534/genetics.119.301906 · Cited by 16
A novel algorithm for simultaneous SNP selection in high-dimensional genome-wide association studiesPreprint
Zuber, V., Pedro, A., Silva, D., & Strimmer, K. (2012) · Cited by 16
Spencer, A. V., Cox, A., & Walters, K. (2013) · Annals of Human Genetics · doi:10.1111/ahg.12043 · Cited by 15
Rejoinder: ‘Gene hunting with hidden Markov model knockoffs’
Sesia, M., Sabatti, C., & Candès, E. J. (2018) · Biometrika · doi:10.1093/biomet/asy075 · Cited by 15
Bayesian Variable Selection in Searching for Additive and Dominant Effects in Genome-Wide Data
Peltola, T., Marttinen, P., Jula, A. M., Salomaa, V. V., Perola, M., & Vehtari, A. (2012) · PLoS ONE · doi:10.1371/journal.pone.0029115 · Cited by 15
Croiseau, P., & Cordell, H. J. (2009) · BMC Proceedings · doi:10.1186/1753-6561-3-s7-s61 · Cited by 15
Searching for improvements in predicting human eye colour from DNA
Kukla‐Bartoszek, M., Teisseyre, P., Pośpiech, E., Karłowska-Pik, J., Zieliński, P., Woźniak, A., Boroń, M., Dąbrowski, M., Zubańska, M., Jarosz, A., et al. (2021) · International Journal of Legal Medicine · doi:10.1007/s00414-021-02645-5 · Cited by 19
BLUPmrMLM: A Fast mrMLM Algorithm in Genome-wide Association Studies
Li, H., Wang, J.-T., Zhao, Q., & Zhang, Y. (2024) · Genomics Proteomics & Bioinformatics · doi:10.1093/gpbjnl/qzae020 · Cited by 14
Haem, E., Harling, K., Ayatollahi, S. M. T., Zare, N., & Karlsson, M. O. (2017) · Journal of Pharmacokinetics and Pharmacodynamics · doi:10.1007/s10928-017-9504-6 · Cited by 14
Identification of Grouped Rare and Common Variants via Penalized Logistic Regression
Ayers, K. L., & Cordell, H. J. (2013) · Genetic Epidemiology · doi:10.1002/gepi.21746 · Cited by 14
Multivariate Methods for Genetic Variants Selection and Risk Prediction in Cardiovascular Diseases
Malovini, A. L., Bellazzi, R., Napolitano, C., & Guffanti, G. (2016) · Frontiers in Cardiovascular Medicine · doi:10.3389/fcvm.2016.00017 · Cited by 13
Pośpiech, E., Karłowska-Pik, J., Kukla‐Bartoszek, M., Woźniak, A., Boroń, M., Zubańska, M., Jarosz, A., Bronikowska, A., Grzybowski, T., Płoski, R., et al. (2022) · Forensic Science International Genetics · doi:10.1016/j.fsigen.2022.102693 · Cited by 13
Population-Based Genomewide Genetic Analysis of Common Clinical Chemistry Analytes
Chasman, D. I., Paré, G., & Ridker, P. M. (2008) · Clinical Chemistry · doi:10.1373/clinchem.2008.107243 · Cited by 13
Bayesian generalized fused lasso modeling via NEG distribution
Shimamura, K., Ueki, M., Kawano, S., & Konishi, S. (2018) · Communication in Statistics- Theory and Methods · doi:10.1080/03610926.2018.1489056 · Cited by 13
Joint genotype‐ and ancestry‐based genome‐wide association studies in admixed populations
Szulc, P., Bogdan, M., Frommlet, F., & Tang, H. (2017) · Genetic Epidemiology · doi:10.1002/gepi.22056 · Cited by 13
Anderson, L. N., Briollais, L., Atkinson, H. C., Marsh, J. A., Xu, J., Connor, K. L., Matthews, S. G., Pennell, C. E., & Lye, S. J. (2014) · PLoS ONE · doi:10.1371/journal.pone.0092957 · Cited by 13
Szefer, E., Lu, D., Nathoo, F. S., Beg, M. F., Graham, J., & Initiative, F. T. A. D. N. (2017) · Statistical Applications in Genetics and Molecular Biology · doi:10.1515/sagmb-2016-0077 · Cited by 11
Genome-Wide Association Studies (GWAS)
de Resende, M. D. V., Silva, F. F. E., Resende, M. F. R., & Azevedo, C. F. (2014) · Elsevier eBooks · doi:10.1016/b978-0-12-418672-9.00004-0 · Cited by 13
Mallick, H., & Tiwari, H. K. (2016) · Frontiers in Genetics · doi:10.3389/fgene.2016.00032 · Cited by 12
Gao, Y., Dong, L., Xu, S., Xiao, S., Fang, M., & Wang, Z. (2017) · Aquaculture · doi:10.1016/j.aquaculture.2017.09.035 · Cited by 12
Using penalized regression to predict phenotype from SNP data
Cherlin, S., Howey, R. A. J., & Cordell, H. J. (2018) · BMC Proceedings · doi:10.1186/s12919-018-0149-2 · Cited by 12
Kang, C., Yu, H., & Yi, G. (2013) · BMC Medical Informatics and Decision Making · doi:10.1186/1472-6947-13-s1-s3 · Cited by 12
Bayesian Variable Selection to identify QTL affecting a simulated quantitative trait
Schurink, A., Janss, L., & Heuven, H. C. M. (2012) · BMC Proceedings · doi:10.1186/1753-6561-6-s2-s8 · Cited by 11
Metadata
- Authors
- Hoggart, C. J., Whittaker, J. C., De Iorio, M., & Balding, D. J.
- Affiliations
- Imperial College London (GB)
- London School of Hygiene & Tropical Medicine (GB)
- Corresponding author
- Hoggart, Clive J.
- Venue
- PLoS Genetics
- Publisher
- Public Library of Science (PLoS)
- Published
- 2008-07-25
- Volume
- 4
- Issue
- 7
- Pages
- e1000130
- ISSN
- 1553-7390 (ISSN-L)
- 1553-7404
- PMC
- PMC2464715
- OpenAlex
- W2095741373
- Cited by
- 321
- References
- 31
- Citations by year
- 2026: 2 · 2025: 5 · 2024: 11 · 2023: 7 · 2022: 8 · 2021: 9 · 2020: 9 · 2019: 20 · 2018: 20 · 2017: 15 · 2016: 16 · 2015: 21 · 2014: 30 · 2013: 29 · 2012: 42
- FWCI
- 14.66
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Locations
- PLoS Genetics (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- LSHTM Research Online (London School of Hygiene and Tropical Medicine) (acceptedVersion)
- Minerva Access (University of Melbourne) (submittedVersion, cc-by)
- RePEc: Research Papers in Economics (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- UCL Discovery (University College London) (submittedVersion)
- Funders
- Medical Research Council
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genetic Associations and Epidemiology
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genetic Mapping and Diversity in Plants and Animals
- Physical Sciences › Mathematics › Statistics and Probability › Statistical Methods and Inference
- Keywords
- genome-wide association study, posterior mode estimation, fine mapping, complex diseases, quantitative phenotypes, gene-gene interaction, stochastic search
- MeSH terms
- Algorithms, Bayes Theorem, Chromosomes, Human, Computer Simulation, Diabetes Mellitus, Type 2, Feasibility Studies, Genotype, Haplotypes, Humans, Software, Linkage Disequilibrium, Genome, Human, Likelihood Functions, Case-Control Studies, Sequence Analysis, DNA, Genetic Predisposition to Disease, Physical Chromosome Mapping, Polymorphism, Single Nucleotide
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
