

The DOI matches its Crossref record; 75 of the 76 references it cites were found in a registry or an index.
A general approach for detecting expressed mutations in AML cells using single cell RNA-sequencing
Open accessAbstract
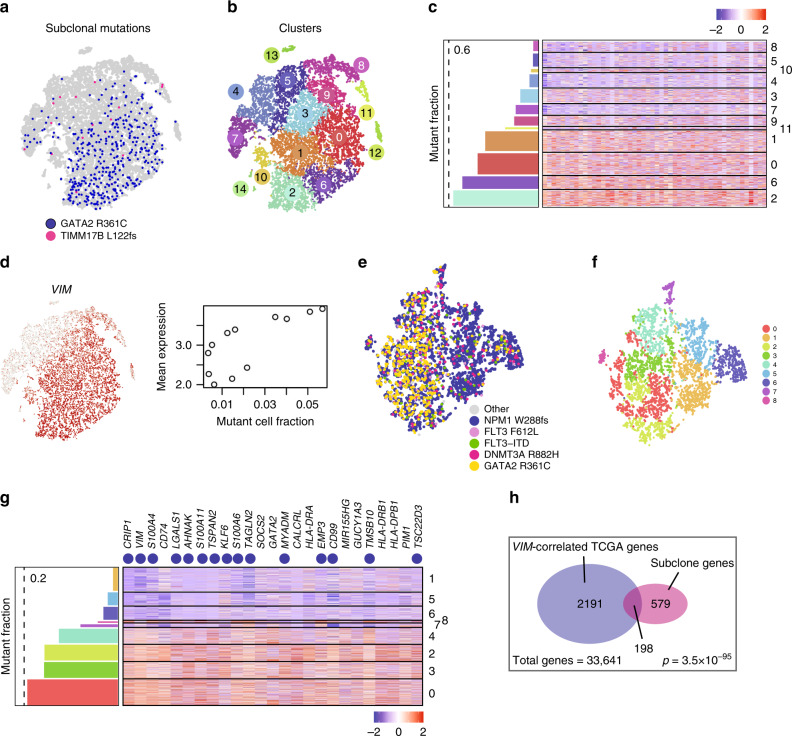
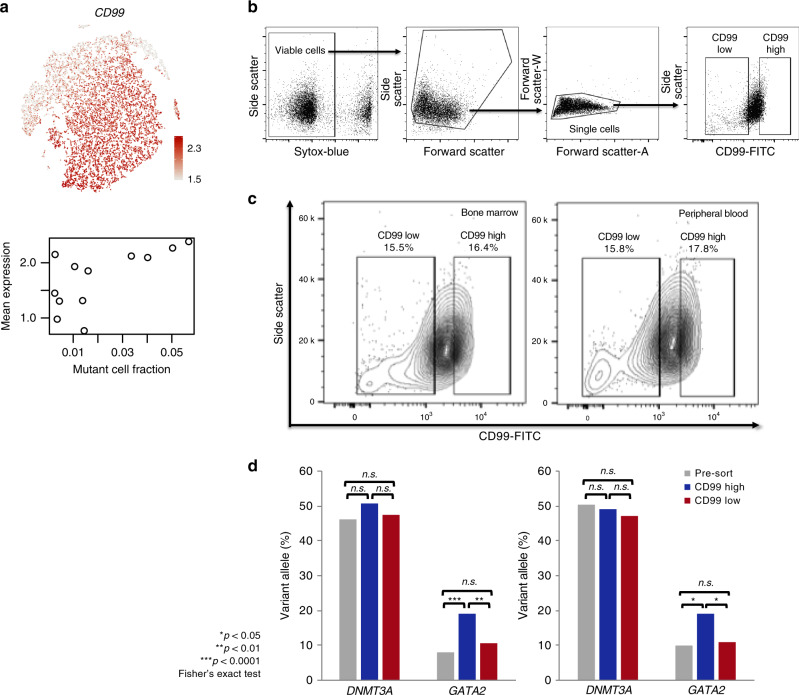
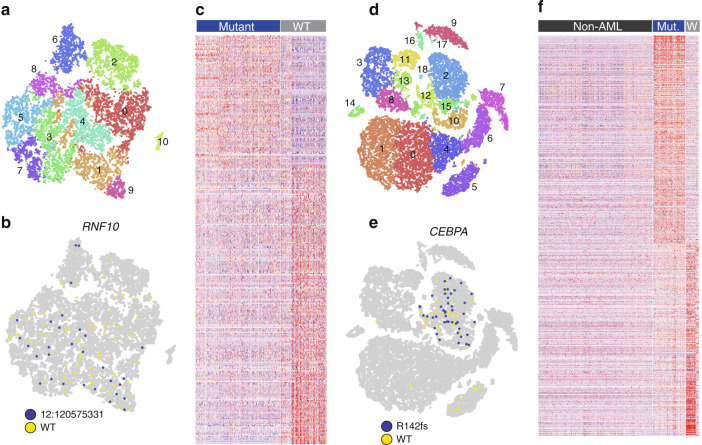
Virtually all tumors are genetically heterogeneous, containing mutationally-defined subclonal cell populations that often have distinct phenotypes. Single-cell RNA-sequencing has revealed that a variety of tumors are also transcriptionally heterogeneous, but the relationship between expression heterogeneity and subclonal architecture is unclear. Here, we address this question in the context of Acute Myeloid Leukemia (AML) by integrating whole genome sequencing with single-cell RNA-sequencing (using the 10x Genomics Chromium Single Cell 5’ Gene Expression workflow). Applying this approach to five cryopreserved AML samples, we identify hundreds to thousands of cells containing tumor-specific mutations in each case, and use the results to distinguish AML cells (including normal-karyotype AML cells) from normal cells, identify expression signatures associated with subclonal mutations, and find cell surface markers that could be used to purify subclones for further study. This integrative approach for connecting genotype to phenotype is broadly applicable to any sample that is phenotypically and genetically heterogeneous.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
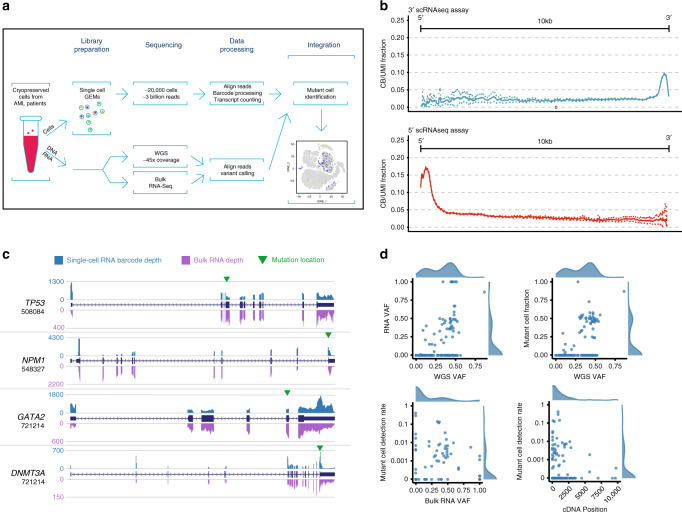
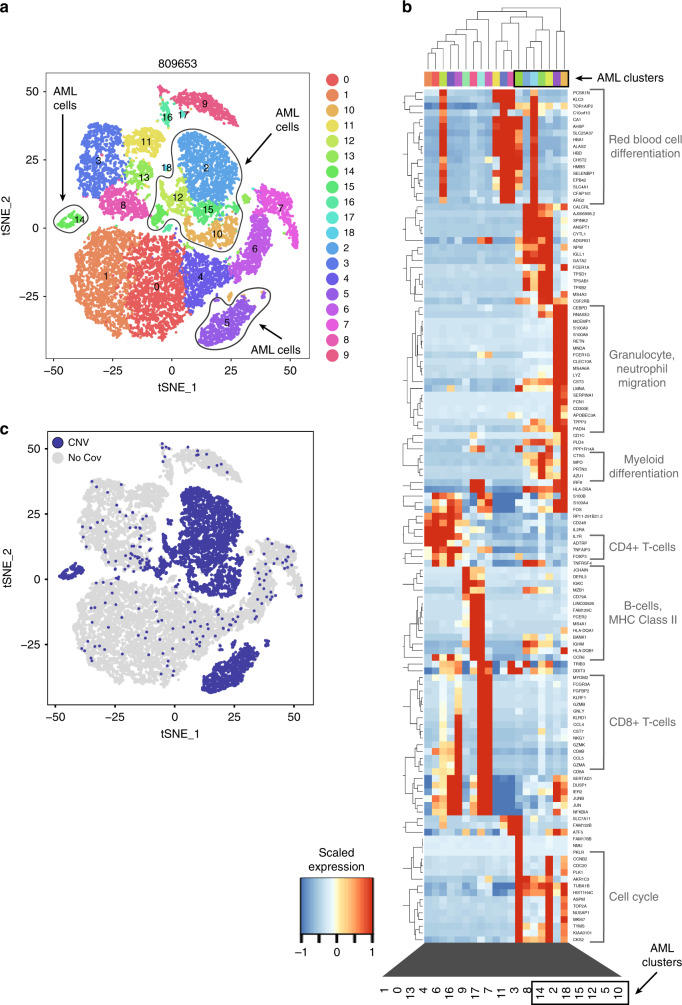
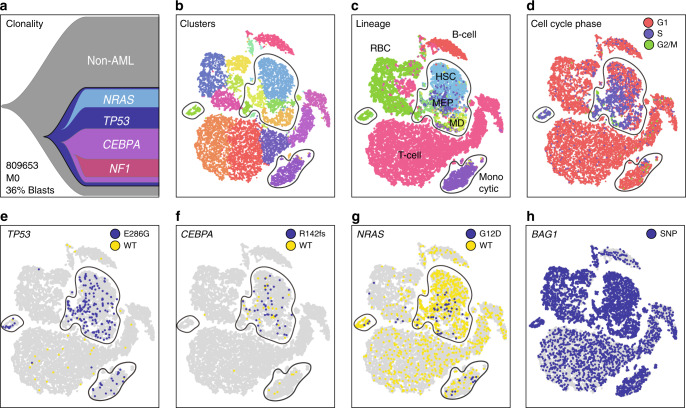
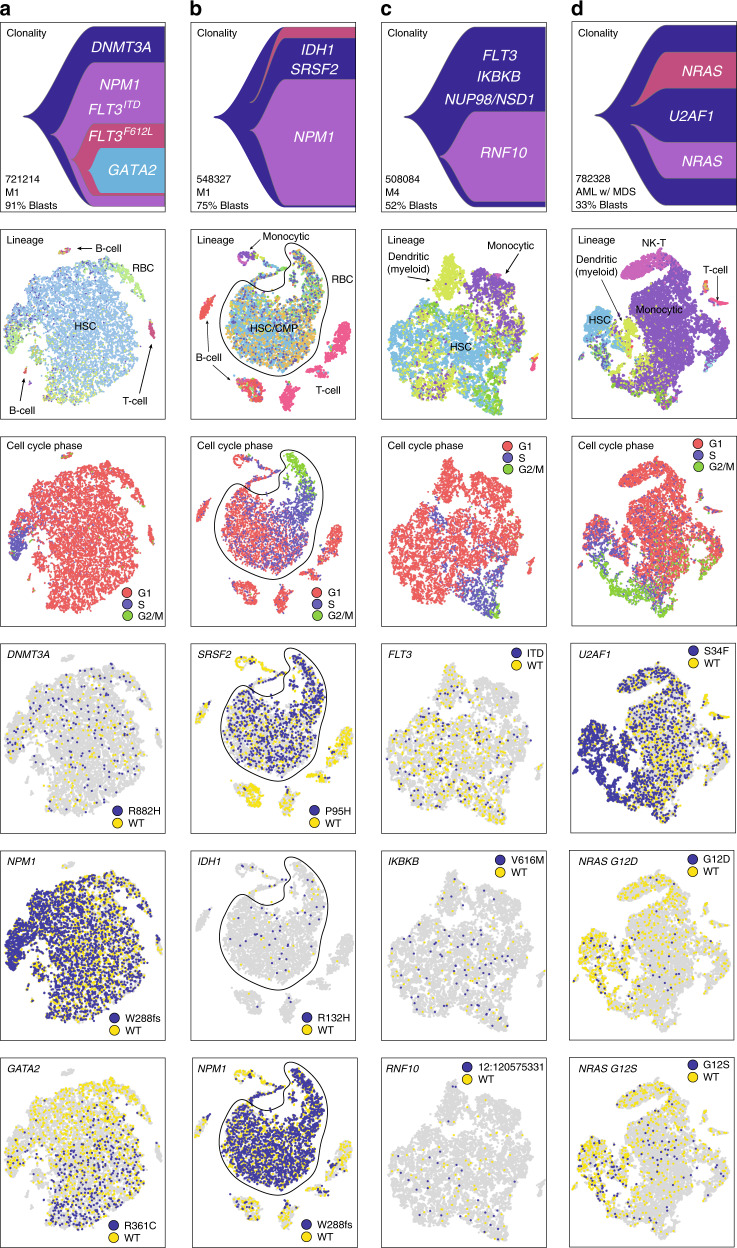
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“Next, we count alleles per cell with vartrix14(Fig. 1c) which avoids reference bias due to ambiguous support such as alignment end effects and corrects bases with duplicate reads via the UMI.”
From Souporcell: robust clustering of single-cell RNA-seq data by genotype without reference genotypes (2020) · cites this paper as [14]
Experimental — accuracy not yet measured
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
The Sequence Alignment/Map format and SAMtools
Li, H., Handsaker, B., Wysoker, A., Fennell, T., Ruan, J., Homer, N., Marth, G., Abecasis, G., Durbin, R., & 1000 Genome Project Data Processing Subgroup (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp352 · Cited by 62,041
Fast and accurate short read alignment with Burrows-Wheeler transform
Li, H., & Durbin, R. (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp324 · Cited by 58,607
The Genome Analysis Toolkit: A MapReduce framework for analyzing next-generation DNA sequencing data
McKenna, A., Hanna, M., Banks, E., Sivachenko, A., Cibulskis, K., Kernytsky, A., Garimella, K., Altshuler, D., Gabriel, S., Daly, M., et al. (2010) · Genome Research · doi:10.1101/gr.107524.110 · Cited by 27,895
Integrating single-cell transcriptomic data across different conditions, technologies, and species
Butler, A., Hoffman, P., Smibert, P., Papalexi, E., & Satija, R. (2018) · Nature Biotechnology · doi:10.1038/nbt.4096 · Cited by 13,174
Comprehensive molecular portraits of human breast tumours
The Cancer Genome Atlas Network (2012) · Nature · doi:10.1038/nature11412 · Cited by 12,129
Fast and accurate long-read alignment with Burrows-Wheeler transform.
Li, H., & Durbin, R. (2010) · Bioinformatics · doi:10.1093/bioinformatics/btp698 · Cited by 12,063
The Human Genome Browser at UCSC
Kent, W. J., Sugnet, C. W., Furey, T. S., Roskin, K. M., Pringle, T. H., Zahler, A. M., & Haussler, A. D. (2002) · Genome Research · doi:10.1101/gr.229102 · Cited by 10,400
Near-optimal probabilistic RNA-seq quantification
Bray, N. L., Pimentel, H., Melsted, P., & Pachter, L. (2016) · Nature Biotechnology · doi:10.1038/nbt.3519 · Cited by 9,723
deepTools2: a next generation web server for deep-sequencing data analysis
Ramírez, F., Ryan, D. P., Grüning, B., Bhardwaj, V., Kilpert, F., Richter, A. S., Heyne, S., Dündar, F., & Manke, T. (2016) · Nucleic Acids Research · doi:10.1093/nar/gkw257 · Cited by 8,096
In situ click chemistry generation of cyclooxygenase-2 inhibitors
Bhardwaj, A., Kaur, J., Wuest, M., & Wuest, F. (2017) · Nature Communications · doi:10.1038/s41467-016-0009-6 · Cited by 7,448
Research works printed but not linked: 1
- The Cancer Genome Atlas Research Network. Genomic and epigenomic landscapes of adult de novo acute myeloid leukemia. N. Engl. J. Med. 368, 2059–2074 (2013). · DOI · Paper page
Cited by
Sorted by citations
Kumar, V., Ramnarayanan, K., Sundar, R., Padmanabhan, N., Srivastava, S., Koiwa, M., Yasuda, T., Koh, V., Huang, K. K., Tay, S. T., et al. (2022) · Cancer Discovery · doi:10.1158/2159-8290.cd-21-0683 · Cited by 573
Souporcell: robust clustering of single-cell RNA-seq data by genotype without reference genotypes
Heaton, H., Talman, A. M., Knights, A., Imaz, M., Gaffney, D. J., Durbin, R., Hemberg, M., & Lawniczak, M. K. N. (2020) · Nature Methods · doi:10.1038/s41592-020-0820-1 · Cited by 506
Single-cell mutation analysis of clonal evolution in myeloid malignancies
Miles, L. A., Bowman, R. L., Merlinsky, T. R., Csete, I., Ooi, A. T., Durruthy-Durruthy, R., Bowman, M., Famulare, C. A., Patel, M. A., Méndez, P., et al. (2020) · Nature · doi:10.1038/s41586-020-2864-x · Cited by 522
Applications of single-cell RNA sequencing in drug discovery and development
de Sande, B. V., Lee, J. S., Mutasa-Gottgens, E., Naughton, B. J., Bacon, W., Manning, J. R., Wang, Y., Pollard, J. R., Mendez, M., Hill, J., et al. (2023) · Nature Reviews Drug Discovery · doi:10.1038/s41573-023-00688-4 · Cited by 462
Integrating genetic and non-genetic determinants of cancer evolution by single-cell multi-omics
Nam, A. S., Chaligné, R., & Landau, D. A. (2020) · Nature Reviews Genetics · doi:10.1038/s41576-020-0265-5 · Cited by 399
Metadata
- Authors
- Petti, A. A., Williams, S. R., Miller, C. A., Fiddes, I. T., Srivatsan, S. N., Chen, D. Y., Fronick, C. C., Fulton, R. S., Church, D. M., & Ley, T. J.
- Affiliations
- Washington University in St. Louis (US)
- 10X Genomics (United States) (US)
- Corresponding author
- Ley, Timothy J.
- ORCID
- Venue
- Nature Communications
- Published
- 2019-08-14
- Volume
- 10
- Issue
- 1
- Pages
- 3660-3660
- ISSN
- 2041-1723 (ISSN-L)
- PMC
- PMC6694122
- OpenAlex
- W2968098426
- Cited by
- 215
- References
- 76
- Citations by year
- 2026: 14 · 2025: 26 · 2024: 35 · 2023: 44 · 2022: 32 · 2021: 40 · 2020: 22 · 2019: 2
- FWCI
- 17.12
- Type
- journal article
- Language
- en
- Update notices
- correction (2022-07-21)
- Open access
- gold
- doi.org
- Locations
- Nature Communications (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Digital Commons@Becker (Washington University School of Medicine) (submittedVersion, cc-by)
- RePEc: Research Papers in Economics (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- Funders
- U.S. Department of Health & Human Services | NIH | National Cancer Institute (CA101937, CA197561, CA167540, CA211782, K12CA167540, R50CA211782)
- Topics
- Health Sciences › Medicine › Hematology › Acute Myeloid Leukemia Research
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Cancer Research › Cancer Genomics and Diagnostics
- Keywords
- acute myeloid leukemia, subclonal heterogeneity, cancer mutations, transcriptional heterogeneity, single-cell RNA sequencing, genotype-phenotype correlation, cell surface markers, normal karyotype AML, tumor cell identification
- MeSH terms
- Whole Genome Sequencing, Genotype, Humans, Mutation, Phenotype, RNA, Leukemia, Myeloid, Acute, Sequence Analysis, RNA, Polymorphism, Single Nucleotide, Gene Expression Profiling, Genomics, Single-Cell Analysis
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
