

The DOI matches its Crossref record; 72 of the 72 references it cites were found in a registry or an index.
SEACells infers transcriptional and epigenomic cellular states from single-cell genomics data
Open accessAbstract
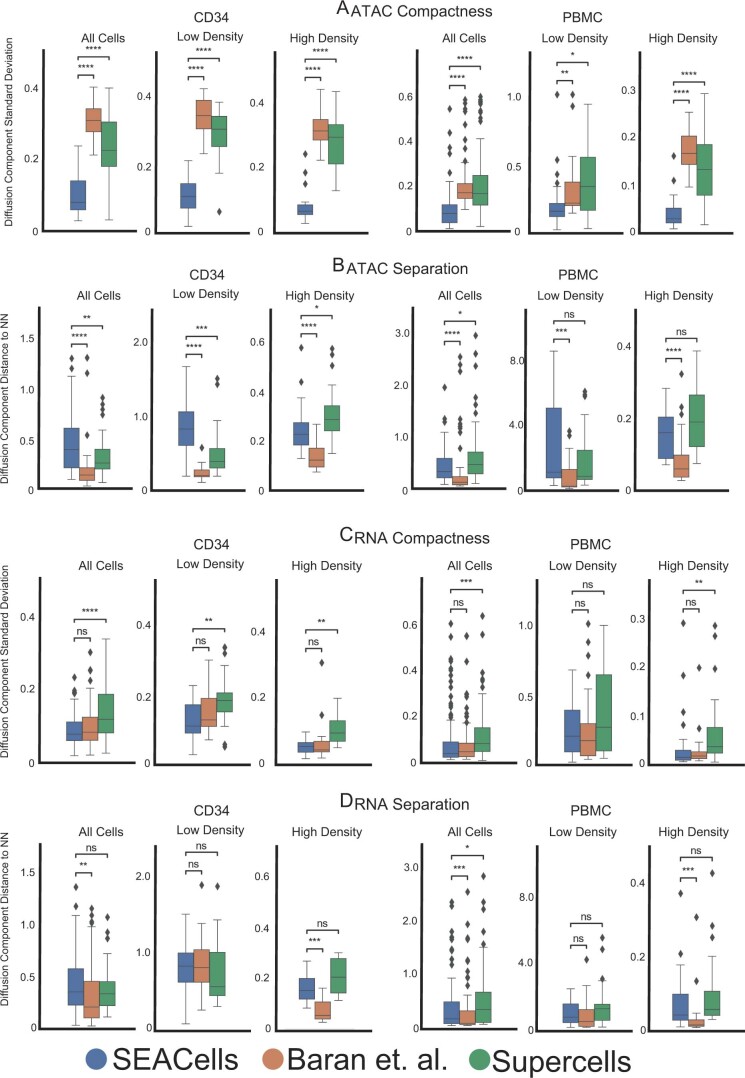
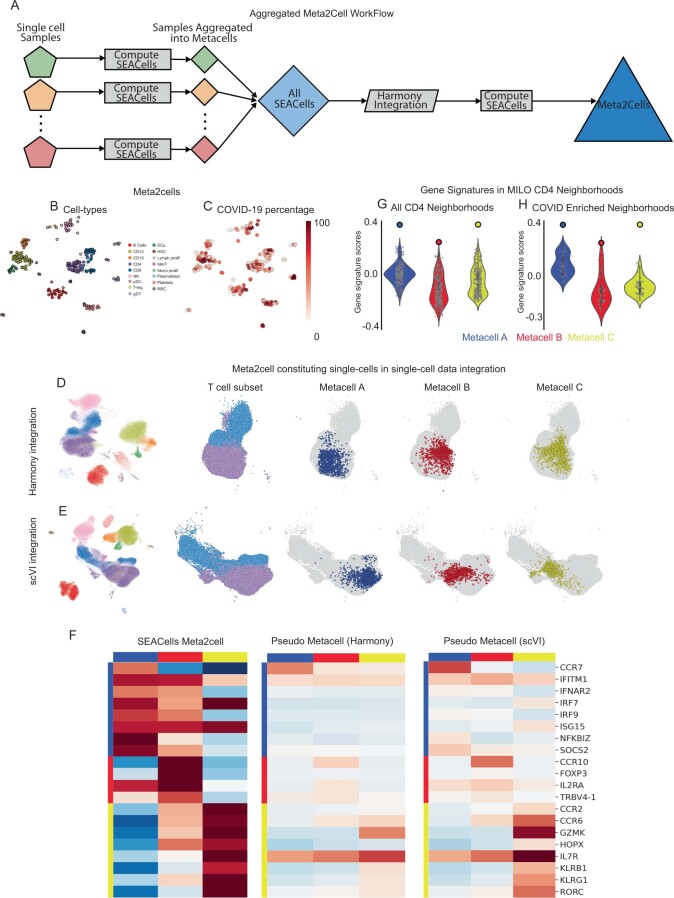
Metacells are cell groupings derived from single-cell sequencing data that represent highly granular, distinct cell states. Here we present single-cell aggregation of cell states (SEACells), an algorithm for identifying metacells that overcome the sparsity of single-cell data while retaining heterogeneity obscured by traditional cell clustering. SEACells outperforms existing algorithms in identifying comprehensive, compact and well-separated metacells in both RNA and assay for transposase-accessible chromatin (ATAC) modalities across datasets with discrete cell types and continuous trajectories. We demonstrate the use of SEACells to improve gene–peak associations, compute ATAC gene scores and infer the activities of critical regulators during differentiation. Metacell-level analysis scales to large datasets and is particularly well suited for patient cohorts, where per-patient aggregation provides more robust units for data integration. We use our metacells to reveal expression dynamics and gradual reconfiguration of the chromatin landscape during hematopoietic differentiation and to uniquely identify CD4 T cell differentiation and activation states associated with disease onset and severity in a Coronavirus Disease 2019 (COVID-19) patient cohort.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
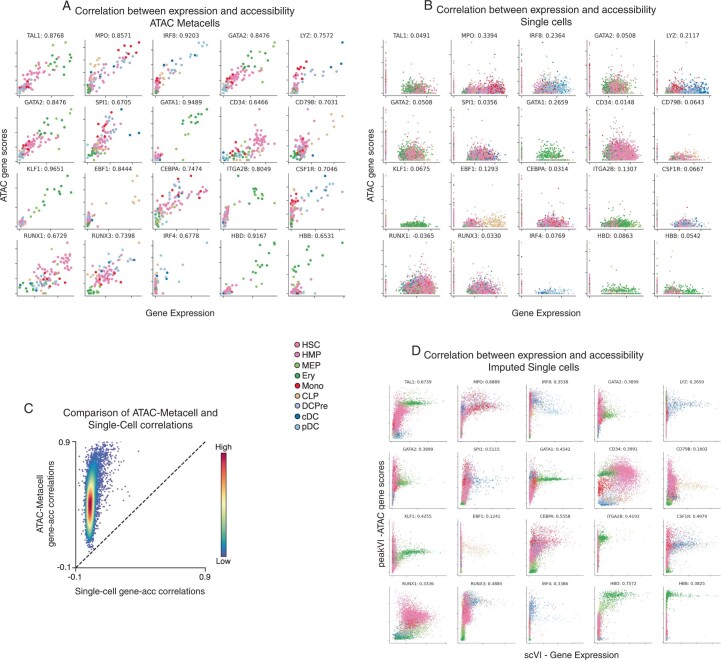
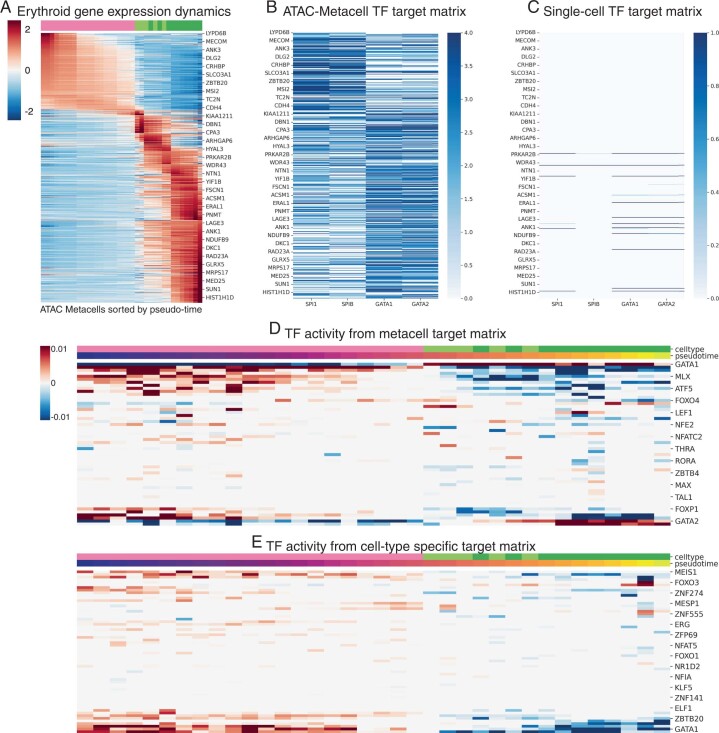
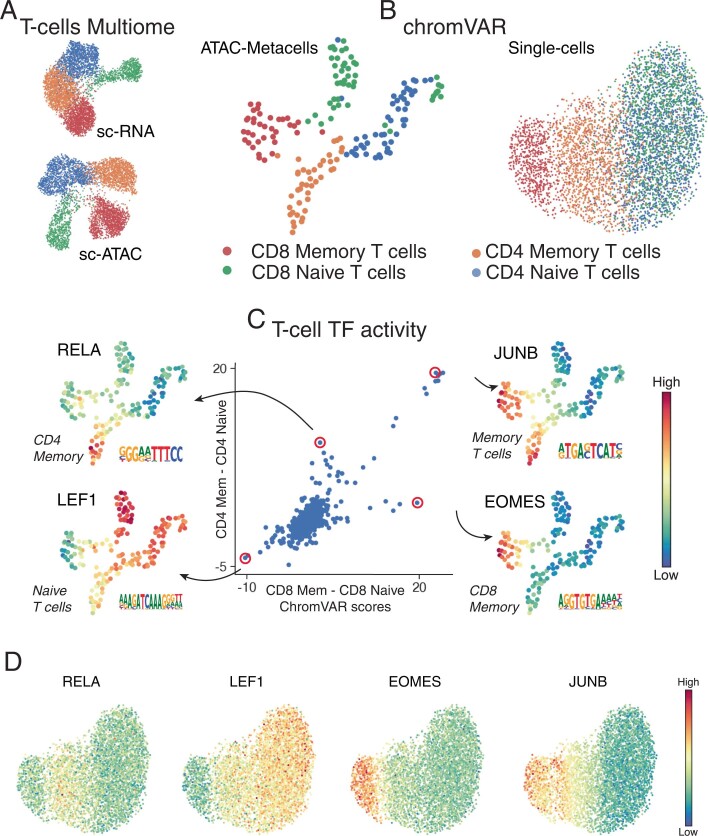
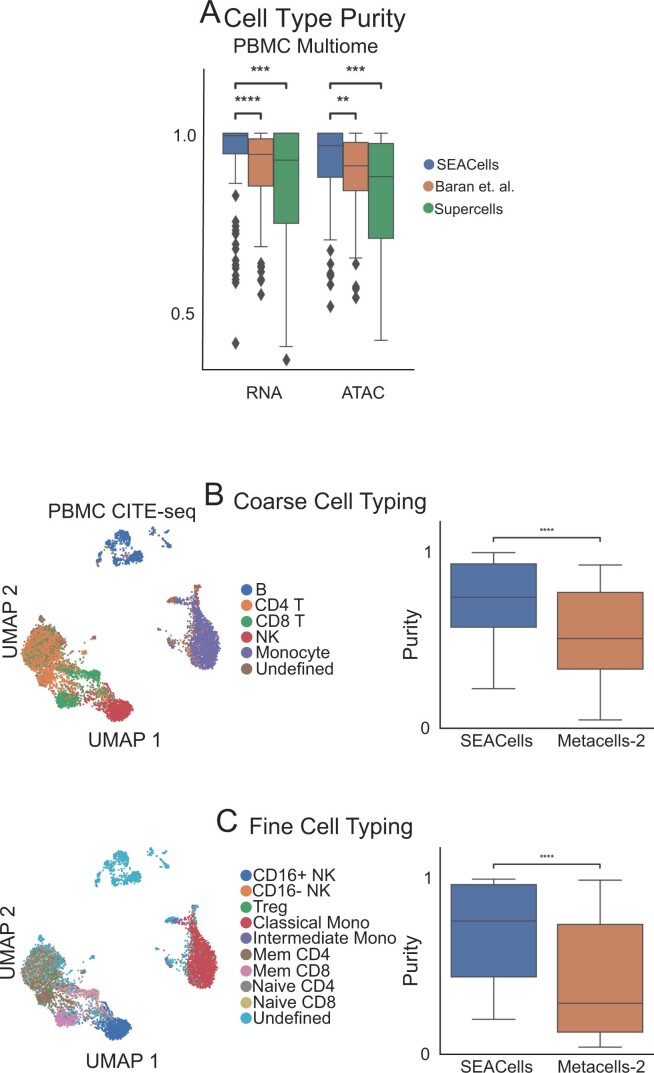
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“We applied hdWGCNA to a dataset of CD34+ hematopoietic stem and progenitor stem cells16 using two additional metacell approaches14,16 and found that all approaches were suitable for downstream network analysis (Figure S2; Data S1).”
From hdWGCNA identifies co-expression networks in high-dimensional transcriptomics data (2023) · cites this paper as [16]
The cited source says what the claim attributes to it.
In the citing paper
“Notably, SEACells and Metacell2 do not account for cell labels in their aggregation procedures, which may result in a number of metacells containing transcriptomes from differently labeled cells.”
From hdWGCNA identifies co-expression networks in high-dimensional transcriptomics data (2023) · cites this paper as [16]
The cited source says what the claim attributes to it.
In the citing paper
“To minimize the influences from stochastic DNA damage signal, we used the SEACell software59 to group the single cells into 300 metacells based on transcriptome profiles.”
From Single-cell parallel analysis of DNA damage and transcriptome reveals selective genome vulnerability (2025) · cites this paper as [59]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Moderated estimation of fold change and dispersion for RNA-seq data with DESeq2
Love, M. I., Huber, W., & Anders, S. (2014) · Genome Biology · doi:10.1186/s13059-014-0550-8 · Cited by 95,106
Subramanian, A., Tamayo, P., Mootha, V. K., Mukherjee, S., Ebert, B. L., Gillette, M. A., Paulovich, A., Pomeroy, S. L., Golub, T. R., Lander, E. S., et al. (2005) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.0506580102 · Cited by 53,625
Regression shrinkage and selection via the lasso
Tibshirani, R. (1996) · Journal of the Royal Statistical Society Series B: Statistical Methodology · doi:10.1111/j.2517-6161.1996.tb02080.x · Cited by 51,482
Model-based Analysis of ChIP-Seq (MACS)
Zhang, Y., Liu, T., Meyer, C. A., Eeckhoute, J., Johnson, D. S., Bernstein, B. E., Nusbaum, C., Myers, R. M., Brown, M., Li, W., et al. (2008) · Genome Biology · doi:10.1186/gb-2008-9-9-r137 · Cited by 18,386
Integrated analysis of multimodal single-cell data
Hao, Y., Hao, S., Andersen-Nissen, E., Mauck, W. M., Zheng, S., Butler, A., Lee, M. J., Wilk, A. J., Darby, C., Zagar, M., et al. (2020) · doi:10.1101/2020.10.12.335331 · Cited by 15,508
Fast, sensitive and accurate integration of single-cell data with Harmony
Korsunsky, I., Millard, N., Fan, J., Slowikowski, K., Zhang, F., Wei, K., Baglaenko, Y., Brenner, M., Loh, P.-R., & Raychaudhuri, S. (2019) · Nature Methods · doi:10.1038/s41592-019-0619-0 · Cited by 10,488
SCANPY: large-scale single-cell gene expression data analysis
Wolf, F. A., Angerer, P., & Theis, F. J. (2018) · Genome Biology · doi:10.1186/s13059-017-1382-0 · Cited by 8,881
Buenrostro, J. D., Giresi, P. G., Zaba, L. C., Chang, H. Y., & Greenleaf, W. J. (2013) · Nature Methods · doi:10.1038/nmeth.2688 · Cited by 6,422
From Louvain to Leiden: guaranteeing well-connected communities
Traag, V. A., Waltman, L., & van Eck, N. J. (2019) · Scientific Reports · doi:10.1038/s41598-019-41695-z · Cited by 5,635
The single-cell transcriptional landscape of mammalian organogenesis
Cao, J., Spielmann, M., Qiu, X., Huang, X., Ibrahim, D. M., Hill, A. J., Zhang, F., Mundlos, S., Christiansen, L., Steemers, F. J., et al. (2019) · Nature · doi:10.1038/s41586-019-0969-x · Cited by 4,490
Research works printed but not linked: 2
- Bauckage, C., Kersting, K., Hoppe, F. & Thurau, C. in Workshop New Challenges in Neural Computation. https://www.techfak.uni-bielefeld.de/~fschleif/mlr/mlr_03_2015.pdf (2015).
- McDaid, A. F., Greene, D. & Hurley, N. Normalized mutual information to evaluate overlapping community finding algorithms. Preprint at https://arxiv.org/abs/1110.2515 (2011). · arXiv
Cited by
Sorted by citations
Dictionary learning for integrative, multimodal and scalable single-cell analysis
Hao, Y., Stuart, T., Kowalski, M. H., Choudhary, S., Hoffman, P., Hartman, A., Srivastava, A., Molla, G., Madad, S., Fernandez-Granda, C., et al. (2024) · Nature Biotechnology · doi:10.1038/s41587-023-01767-y · Cited by 5,631
hdWGCNA identifies co-expression networks in high-dimensional transcriptomics data
Morabito, S., Reese, F., Rahimzadeh, N., Miyoshi, E., & Swarup, V. (2023) · Cell Reports Methods · doi:10.1016/j.crmeth.2023.100498 · Cited by 773
Gene regulatory network inference in the era of single-cell multi-omics
Badia-i-Mompel, P., Wessels, L., Müller‐Dott, S., Trimbour, R., Flores, R. O. R., Argelaguet, R., & Sáez-Rodríguez, J. (2023) · Nature Reviews Genetics · doi:10.1038/s41576-023-00618-5 · Cited by 507
Lineage plasticity in prostate cancer depends on JAK/STAT inflammatory signaling
Chan, J. M., Zaidi, S., Love, J. R., Zhao, J. L., Setty, M. N., Wadosky, K. M., Gopalan, A., Choo, Z.-N., Persad, S., Choi, J., et al. (2022) · Science · doi:10.1126/science.abn0478 · Cited by 359
Self-citationProfiling cell identity and tissue architecture with single-cell and spatial transcriptomics
Gulati, G. S., D'Silva, J. P., Liu, Y., Wang, L., & Newman, A. M. (2024) · Nature Reviews Molecular Cell Biology · doi:10.1038/s41580-024-00768-2 · Cited by 347
Metadata
- Authors
- Persad, S., Choo, Z.-N., Dien, C., Sohail, N., Masilionis, I., Chaligné, R., Nawy, T., Brown, C. C., Sharma, R., Pe’er, I., Setty, M., & Pe’er, D.
- Affiliations
- Memorial Sloan Kettering Cancer Center (US)
- Fred Hutch Cancer Center (US)
- Columbia University (US)
- Corresponding author
- Pe’er, Itsik
- Setty, Manu
- Pe’er, Dana
- ORCID
- Venue
- Nature Biotechnology
- Published
- 2023-03-27
- Volume
- 41
- Issue
- 12
- Pages
- 1746-1757
- ISSN
- 0733-222X (ISSN-L)
- 1087-0156
- 1546-1696
- PMC
- PMC10713451
- OpenAlex
- W4360999291
- Cited by
- 222
- References
- 72
- Citations by year
- 2026: 72 · 2025: 54 · 2024: 72 · 2023: 20
- FWCI
- 19.87
- Type
- journal article
- Language
- en
- Open access
- hybrid
- doi.org
- Locations
- Nature Biotechnology (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Funders
- U.S. Department of Health & Human Services | NIH | National Institute of General Medical Sciences (R35 GM147125)
- U.S. Department of Health & Human Services | NIH | National Cancer Institute (U54 CA209975, U2C CA233284)
- Howard Hughes Medical Institute
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene Regulatory Network Analysis
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Chromatin Dynamics
- Keywords
- metacells, transcriptional heterogeneity, chromatin accessibility, ATAC-seq, single-cell RNA sequencing, hematopoietic differentiation, CD4+ T cell activation, COVID-19 pandemic, data integration
- MeSH terms
- Algorithms, Chromatin, Humans, CD4-Positive T-Lymphocytes, Genomics, Epigenomics, Single-Cell Analysis
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
