

The DOI matches its Crossref record; 32 of the 32 references it cites were found in a registry or an index.
Batch effect correction for genome-wide methylation data with Illumina Infinium platform
Open accessAbstract
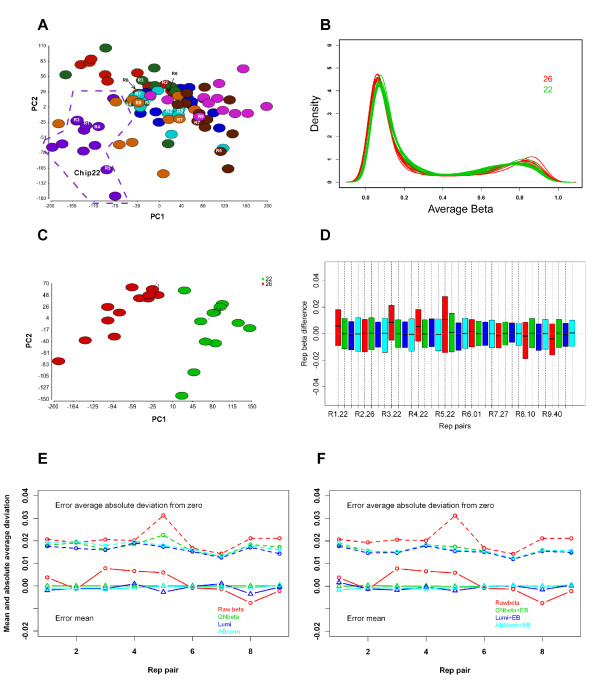
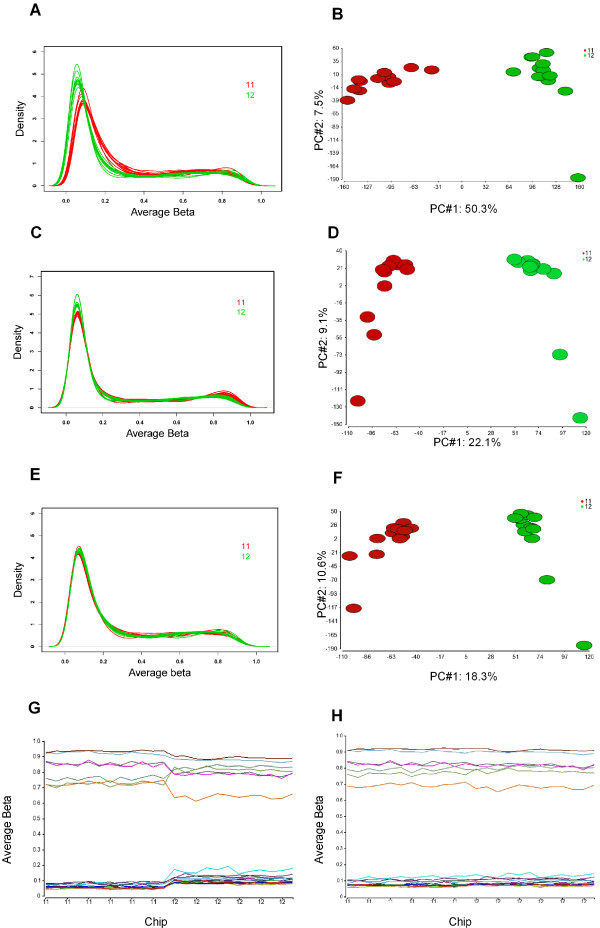
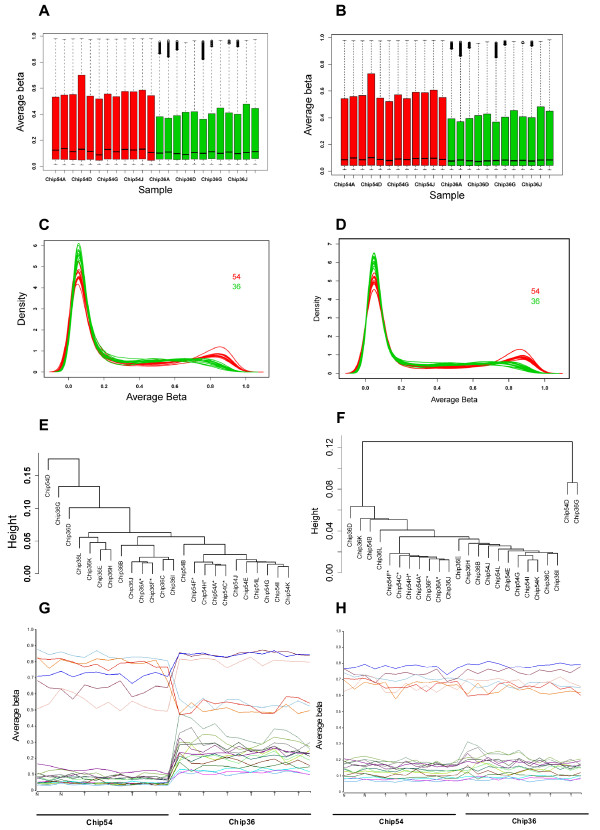
BACKGROUND: Genome-wide methylation profiling has led to more comprehensive insights into gene regulation mechanisms and potential therapeutic targets. Illumina Human Methylation BeadChip is one of the most commonly used genome-wide methylation platforms. Similar to other microarray experiments, methylation data is susceptible to various technical artifacts, particularly batch effects. To date, little attention has been given to issues related to normalization and batch effect correction for this kind of data. METHODS: We evaluated three common normalization approaches and investigated their performance in batch effect removal using three datasets with different degrees of batch effects generated from HumanMethylation27 platform: quantile normalization at average β value (QNβ); two step quantile normalization at probe signals implemented in "lumi" package of R (lumi); and quantile normalization of A and B signal separately (ABnorm). Subsequent Empirical Bayes (EB) batch adjustment was also evaluated. RESULTS: Each normalization could remove a portion of batch effects and their effectiveness differed depending on the severity of batch effects in a dataset. For the dataset with minor batch effects (Dataset 1), normalization alone appeared adequate and "lumi" showed the best performance. However, all methods left substantial batch effects intact in the datasets with obvious batch effects and further correction was necessary. Without any correction, 50 and 66 percent of CpGs were associated with batch effects in Dataset 2 and 3, respectively. After QNβ, lumi or ABnorm, the number of CpGs associated with batch effects were reduced to 24, 32, and 26 percent for Dataset 2; and 37, 46, and 35 percent for Dataset 3, respectively. Additional EB correction effectively removed such remaining non-biological effects. More importantly, the two-step procedure almost tripled the numbers of CpGs associated with the outcome of interest for the two datasets. CONCLUSION: Genome-wide methylation data from Infinium Methylation BeadChip can be susceptible to batch effects with profound impacts on downstream analyses and conclusions. Normalization can reduce part but not all batch effects. EB correction along with normalization is recommended for effective batch effect removal.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“the disappearance of batch effects is a useful indicator [12], but not sufficient as a performance indicator because it does not indicate whether true differences can”
From A data-driven approach to preprocessing Illumina 450K methylation array data (2013) · cites this paper as [12]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Adjusting batch effects in microarray expression data using empirical Bayes methods
Johnson, W. E., Li, C., & Rabinovic, A. (2007) · Biostatistics · doi:10.1093/biostatistics/kxj037 · Cited by 8,658
Bolstad, B. M., Irizarry, R. A., Åstrand, M., & Speed, T. P. (2003) · Bioinformatics · doi:10.1093/bioinformatics/19.2.185 · Cited by 8,328
Tackling the widespread and critical impact of batch effects in high-throughput data
Leek, J. T., Scharpf, R. B., Bravo, H. C., Simcha, D., Langmead, B., Johnson, W. E., Geman, D., Baggerly, K., & Irizarry, R. A. (2010) · Nature Reviews Genetics · doi:10.1038/nrg2825 · Cited by 2,196
lumi : a pipeline for processing Illumina microarray
Du, P., Kibbe, W. A., & Lin, S. M. (2008) · Bioinformatics · doi:10.1093/bioinformatics/btn224 · Cited by 2,028
Singular value decomposition for genome-wide expression data processing and modeling
Alter, O., Brown, P. O., & Botstein, D. (2000) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.97.18.10101 · Cited by 1,989
Capturing Heterogeneity in Gene Expression Studies by Surrogate Variable Analysis
Leek, J. T., & Storey, J. D. (2007) · PLoS Genetics · doi:10.1371/journal.pgen.0030161 · Cited by 1,930
Principles and challenges of genome-wide DNA methylation analysis
Laird, P. W. (2010) · Nature Reviews Genetics · doi:10.1038/nrg2732 · Cited by 1,611
The power and the promise of DNA methylation markers
Laird, P. W. (2003) · Nature reviews. Cancer · doi:10.1038/nrc1045 · Cited by 1,573
Impaired hydroxylation of 5-methylcytosine in myeloid cancers with mutant TET2
Ko, M., Huang, Y., Jankowska, A. M., Pape, U. J., Tahiliani, M., Bandukwala, H. S., An, J., Lamperti, E. D., Koh, K. P., Ganetzky, R. D., et al. (2010) · Nature · doi:10.1038/nature09586 · Cited by 1,340
Age-dependent DNA methylation of genes that are suppressed in stem cells is a hallmark of cancer
Teschendorff, A. E., Menon, U., Gentry-Maharaj, A., Ramus, S. J., Weisenberger, D. J., Shen, H., Campan, M., Noushmehr, H., Bell, C. G., Maxwell, A. P., et al. (2010) · Genome Research · doi:10.1101/gr.103606.109 · Cited by 855
Cited by
Sorted by citations
Items 101–107 of 107
Pontali, G., Cascione, L., Arribas, A. J., Rinaldi, A., Bertoni, F., & Giugno, R. (2019) · Lecture notes in computer science · doi:10.1007/978-3-030-17935-9_3
Quantification Methods for Methylation Levels in Illumina Arrays
Ryu, D., & Shen, H. (2012) · Methods in molecular biology · doi:10.1007/978-1-0716-1994-0_1
Odintsova, V. V., Willemsen, G. A. H. M., Dolan, C. V., Hottenga, J., Martin, N. G., Slagboom, P. E., Ordoñana, J. R., & Boomsma, D. I. (2022) · Elsevier eBooks · doi:10.1016/b978-0-12-821514-2.00011-8
Desenvolvimento de um pipeline para anáise genômica de metilação do DNA
Vieira, H. C. (2016) · Digital Library of Theses and Dissertations (Universidade de São Paulo) · doi:10.11606/d.95.2016.tde-20230725-114652
Mizuno, T., & Kusuhara, H. (2024) · The Journal of Toxicological Sciences · doi:10.2131/jts.49.249
Tyagi, S. (2026) · Proceedings of the Indian National Science Academy · doi:10.1007/s43538-026-00766-2 · Cited by 1
Metadata
- Authors
- Sun, Z., Chai, H. S., Wu, Y., White, W. M., Donkena, K. V., Klein, C. J., Garovic, V. D., Therneau, T. M., & Kocher, J.-P. A.
- Affiliations
- Mayo Clinic (US)
- Corresponding author
- Kocher, Jean-Pierre A
- ORCID
- Venue
- BMC Medical Genomics
- Published
- 2011-12
- Volume
- 4
- Issue
- 1
- Pages
- 84-84
- ISSN
- 1755-8794 (ISSN-L)
- PMC
- PMC3265417
- OpenAlex
- W2108947463
- Cited by
- 108
- References
- 32
- Citations by year
- 2026: 1 · 2025: 2 · 2024: 4 · 2023: 1 · 2022: 6 · 2021: 4 · 2020: 3 · 2019: 9 · 2018: 9 · 2017: 6 · 2016: 13 · 2015: 13 · 2014: 11 · 2013: 17 · 2012: 9
- FWCI
- 4.81
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- Locations
- BMC Medical Genomics (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- Funders
- Mayo Clinic
- Mayo Foundation for Medical Education and Research
- Center for Individualized Medicine, Mayo Clinic
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Epigenetics and DNA Methylation
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene expression and cancer classification
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Keywords
- batch effect correction, DNA methylation, genome-wide methylation profiling, quantile normalization, data normalization, CpG methylation
- MeSH terms
- Adult, Humans, Male, Reproducibility of Results, Genome, Human, CpG Islands, DNA Methylation, Oligonucleotide Array Sequence Analysis, Databases, Genetic
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
