

The DOI matches its Crossref record; 29 of the 29 references it cites were found in a registry or an index.
Ultrafast and memory-efficient alignment of short DNA sequences to the human genome.
Open accessAbstract
Bowtie is an ultrafast, memory-efficient alignment program for aligning short DNA sequence reads to large genomes. For the human genome, Burrows-Wheeler indexing allows Bowtie to align more than 25 million reads per CPU hour with a memory footprint of approximately 1.3 gigabytes. Bowtie extends previous Burrows-Wheeler techniques with a novel quality-aware backtracking algorithm that permits mismatches. Multiple processor cores can be used simultaneously to achieve even greater alignment speeds. Bowtie is open source (http://bowtie.cbcb.umd.edu).
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
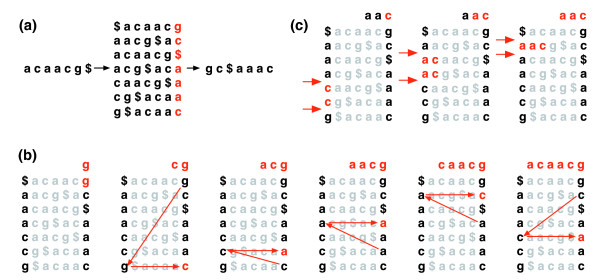
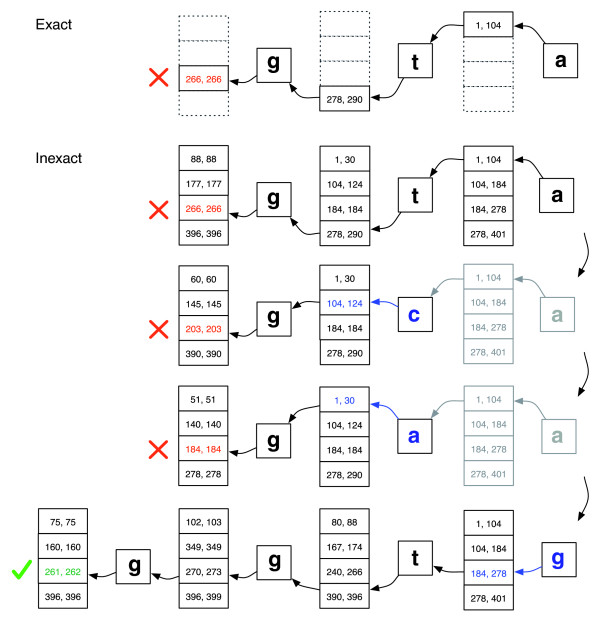
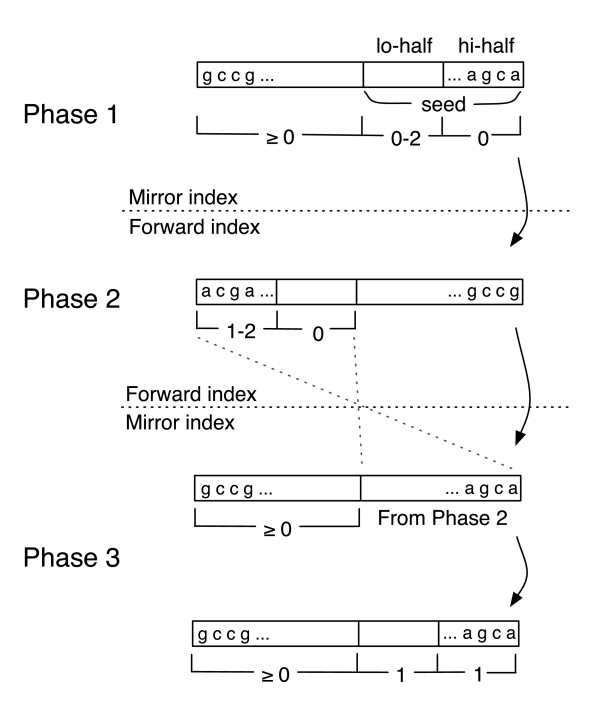
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“To further assess and/or visualize the assembly, we mapped the reads onto the final assembled transcripts using the Bowtie program [27] available at the Galaxy website (http://main.g2.bx.psu.edu/) [23]–[25].”
From Digital Gene Expression Analysis Based on Integrated De Novo Transcriptome Assembly of Sweet Potato [Ipomoea batatas (L.) Lam.] (2012) · cites this paper as [27]
The cited source says what the claim attributes to it.
In the citing paper
“Mapping reads to specific genes is relatively straightforward with the bioinformatics tools available today [10-13], but using these tools to distinguish between reads derived from alternative alleles of the same gene remains challenging [9].”
From Sources of bias in measures of allele-specific expression derived from RNA-seq data aligned to a single reference genome (2013) · cites this paper as [10]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Identification of common molecular subsequences
Smith, T. F., & Waterman, M. S. (1981) · Journal of Molecular Biology · doi:10.1016/0022-2836(81)90087-5 · Cited by 9,962
Base-Calling of Automated Sequencer Traces Using Phred. I. Accuracy AssessmentPreprint
Ewing, B., Hillier, L., Wendl, M. C., & Green, P. (1998) · Genome Research · doi:10.1101/gr.8.3.175 · Cited by 6,993
Base-Calling of Automated Sequencer Traces Using Phred. II. Error Probabilities
Ewing, B., & Green, P. (1998) · Genome Research · doi:10.1101/gr.8.3.186 · Cited by 5,439
Accurate whole human genome sequencing using reversible terminator chemistry
Bentley, D. R., Balasubramanian, S., Swerdlow, H. P., Smith, G. P., Milton, J., Brown, C. G., Hall, K. P., Evers, D. J., Barnes, C. L., Bignell, H. R., et al. (2008) · Nature · doi:10.1038/nature07517 · Cited by 3,684
SOAP: Short oligonucleotide alignment program
Li, R., Li, Y., Kristiansen, K., & Wang, J. (2008) · Bioinformatics · doi:10.1093/bioinformatics/btn025 · Cited by 3,597
Genome-Wide Mapping of in Vivo Protein-DNA Interactions
Johnson, D. S., Mortazavi, A., Myers, R. M., & Wold, B. (2007) · Science · doi:10.1126/science.1141319 · Cited by 2,927
RNA-seq: An assessment of technical reproducibility and comparison with gene expression arrays
Marioni, J. C., Mason, C. E., Mane, S. M., Stephens, M., & Gilad, Y. (2008) · Genome Research · doi:10.1101/gr.079558.108 · Cited by 2,771
Mapping short DNA sequencing reads and calling variants using mapping quality scores
Li, H., Ruan, J., & Durbin, R. (2008) · Genome Research · doi:10.1101/gr.078212.108 · Cited by 2,638
The Transcriptional Landscape of the Yeast Genome Defined by RNA Sequencing
Nagalakshmi, U., Wang, Z., Waern, K., Shou, C., Raha, D., Gerstein, M., & Snyder, M. (2008) · Science · doi:10.1126/science.1158441 · Cited by 2,479
A Block-sorting Lossless Data Compression Algorithm
Burrows, M. T., & Wheeler, D. J. (1994) · Cited by 2,333
DNA sequencing of a cytogenetically normal acute myeloid leukaemia genome
Ley, T. J., Mardis, E. R., Ding, L., Fulton, B., McLellan, M. D., Chen, K., Dooling, D. J., Dunford-Shore, B. H., McGrath, S. D., Hickenbotham, M. T., et al. (2008) · Nature · doi:10.1038/nature07485 · Cited by 1,434
Opportunistic data structures with applications
Ferragina, P., & Manzini, G. (2000) · Proceedings 41st Annual Symposium on Foundations of Computer Science · doi:10.1109/sfcs.2000.892127 · Cited by 1,325
The diploid genome sequence of an Asian individual
Wang, J., Wang, W., Li, R., Li, Y., Tian, G., Goodman, L., Fan, W., Zhang, J., Li, J., Zhang, J., et al. (2008) · Nature · doi:10.1038/nature07484 · Cited by 924
PatternHunter: faster and more sensitive homology search
Ma, B., Tromp, J., & Li, M. (2002) · Bioinformatics · doi:10.1093/bioinformatics/18.3.440 · Cited by 837
Campbell, P. J., Stephens, P. J., Pleasance, E., O’Meara, S., Li, H., Santarius, T., Stebbings, L. A., Leroy, C., Edkins, S., Hardy, C. W., et al. (2008) · Nature Genetics · doi:10.1038/ng.128 · Cited by 827
A Bayesian deconvolution strategy for immunoprecipitation-based DNA methylome analysis
Down, T. A., Rakyan, V. K., Turner, D., Flicek, P., Li, H., Kulesha, E., Gräf, S., Johnson, N., Herrero, J., Tomazou, E. M., et al. (2008) · Nature Biotechnology · doi:10.1038/nbt1414 · Cited by 650
High-throughput sequencing provides insights into genome variation and evolution in Salmonella Typhi
Holt, K. E., Parkhill, J., Mazzoni, C. J., Roumagnac, P., Weill, F., Goodhead, I., Rance, R., Baker, S. G., Maskell, D. J., Wain, J. R., et al. (2008) · Nature Genetics · doi:10.1038/ng.195 · Cited by 498
SeqAn An efficient, generic C++ library for sequence analysis
Gogol‐Döring, A., Weese, D., Rausch, T., & Reinert, K. (2008) · BMC Bioinformatics · doi:10.1186/1471-2105-9-11 · Cited by 338
Using quality scores and longer reads improves accuracy of Solexa read mapping
Smith, A. D., Xuan, Z., & Zhang, M. Q. (2008) · BMC Bioinformatics · doi:10.1186/1471-2105-9-128 · Cited by 336
ZOOM! Zillions of oligos mapped
Lin, H., Zhang, Z., Zhang, M. Q., Ma, B., & Li, M. (2008) · Bioinformatics · doi:10.1093/bioinformatics/btn416 · Cited by 210
An experimental study of an opportunistic index
Ferragina, P., & Manzini, G. (2001) · doi:10.5555/365411.365458 · Cited by 166
Compressed indexing and local alignment of DNA
Lam, T.-W., Sung, W.-K., Tam, S.-L., Wong, C., & Yiu, S.-M. (2008) · Bioinform. · doi:10.1093/bioinformatics/btn032 · Cited by 159
Better Filtering with Gapped q-Grams
Burkhardt, S., & Kärkkäinen, J. (2001) · Lecture notes in computer science · doi:10.1007/3-540-48194-x_6 · Cited by 143
A Plan to Capture Human Diversity in 1000 Genomes
Kaiser, J. (2008) · Science · doi:10.1126/science.319.5862.395 · Cited by 131
Fast BWT in small space by blockwise suffix sorting
Kärkkäinen, J. (2007) · Theoretical Computer Science · doi:10.1016/j.tcs.2007.07.018 · Cited by 91
Fast and practical approximate string matching
Baeza-Yates, R. A., & Perleberg, C. H. (1996) · Information Processing Letters · doi:10.1016/0020-0190(96)00083-x · Cited by 87
Annotating Large Genomes With Exact Word MatchesPreprint
Healy, J., Thomas, E. E., Schwartz, J. T., & Wigler, M. H. (2003) · Genome Research · doi:10.1101/gr.1350803 · Cited by 70
Optimized design and assessment of whole genome tiling arrays
Gräf, S., Nielsen, F. G. G., Kurtz, S., Huynen, M. A., Birney, E., Stunnenberg, H. G., & Flicek, P. (2007) · Bioinformatics · doi:10.1093/bioinformatics/btm200 · Cited by 64
Space-Efficient Whole Genome Comparisons with Burrows–Wheeler Transforms
Lippert, R. A. (2005) · Journal of Computational Biology · doi:10.1089/cmb.2005.12.407 · Cited by 46
Cited by
Sorted by citations
Fast and accurate short read alignment with Burrows-Wheeler transform
Li, H., & Durbin, R. (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp324 · Cited by 58,607
The Sequence Alignment/Map format and SAMtools
Li, H., Handsaker, B., Wysoker, A., Fennell, T., Ruan, J., Homer, N., Marth, G., Abecasis, G., Durbin, R., & 1000 Genome Project Data Processing Subgroup (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp352 · Cited by 62,041
Fast gapped-read alignment with Bowtie 2
Langmead, B., & Salzberg, S. L. (2012) · Nature Methods · doi:10.1038/nmeth.1923 · Cited by 56,400
Self-citationfeatureCounts: an efficient general purpose program for assigning sequence reads to genomic features
Liao, Y., Smyth, G. K., & Shi, W. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btt656 · Cited by 27,219
RSEM: accurate transcript quantification from RNA-Seq data with or without a reference genome
Li, B., & Dewey, C. N. (2011) · BMC Bioinformatics · doi:10.1186/1471-2105-12-323 · Cited by 22,179
Metadata
- Authors
- Langmead, B., Trapnell, C., Pop, M., & Salzberg, S. L.
- Affiliations
- University of Maryland, College Park (US)
- Corresponding author
- Langmead, Ben
- Venue
- Genome Biology
- Published
- 2009-03-04
- Volume
- 10
- Issue
- 3
- Pages
- R25-R25
- ISSN
- 1465-6906 (ISSN-L)
- 1474-760X
- PMC
- PMC2690996
- OpenAlex
- W2124985265
- Cited by
- 21,849
- References
- 29
- Citations by year
- 2026: 620 · 2025: 981 · 2024: 1140 · 2023: 1226 · 2022: 1405 · 2021: 1665 · 2020: 1652 · 2019: 1726 · 2018: 1653 · 2017: 1702 · 2016: 1538 · 2015: 1620 · 2014: 1565 · 2013: 1325 · 2012: 1029
- FWCI
- 267.58
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Locations
- Genome biology (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- University Libraries (University of Maryland) (submittedVersion)
- Digital Repository at the University of Maryland (University of Maryland College Park)
- Funders
- National Institutes of Health
- National Human Genome Research Institute (R01HG004885)
- National Institute of General Medical Sciences (R01GM083873)
- U.S. National Library of Medicine (R01LM006845)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Physical Sciences › Mathematics › Theoretical Computer Science › Algorithms and Data Compression
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Fractal and DNA sequence analysis
- Keywords
- Bowtie, human genome, memory-efficient alignment, parallel alignment, next-generation sequencing reads
- MeSH terms
- Algorithms, Base Sequence, Humans, Genome, Human, Sequence Alignment
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
