

The DOI matches its Crossref record; 31 of the 33 references it cites were found in a registry or an index.
Analysis of the prostate cancer cell line LNCaP transcriptome using a sequencing-by-synthesis approach
Open accessAbstract
BACKGROUND: High throughput sequencing-by-synthesis is an emerging technology that allows the rapid production of millions of bases of data. Although the sequence reads are short, they can readily be used for re-sequencing. By re-sequencing the mRNA products of a cell, one may rapidly discover polymorphisms and splice variants particular to that cell. RESULTS: We present the utility of massively parallel sequencing by synthesis for profiling the transcriptome of a human prostate cancer cell-line, LNCaP, that has been treated with the synthetic androgen, R1881. Through the generation of approximately 20 megabases (MB) of EST data, we detect transcription from over 10,000 gene loci, 25 previously undescribed alternative splicing events involving known exons, and over 1,500 high quality single nucleotide discrepancies with the reference human sequence. Further, we map nearly 10,000 ESTs to positions on the genome where no transcription is currently predicted to occur. We also characterize various obstacles with using sequencing by synthesis for transcriptome analysis and propose solutions to these problems. CONCLUSION: The use of high-throughput sequencing-by-synthesis methods for transcript profiling allows the specific and sensitive detection of many of a cell's transcripts, and also allows the discovery of high quality base discrepancies, and alternative splice variants. Thus, this technology may provide an effective means of understanding various disease states, discovering novel targets for disease treatment, and discovery of novel transcripts.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
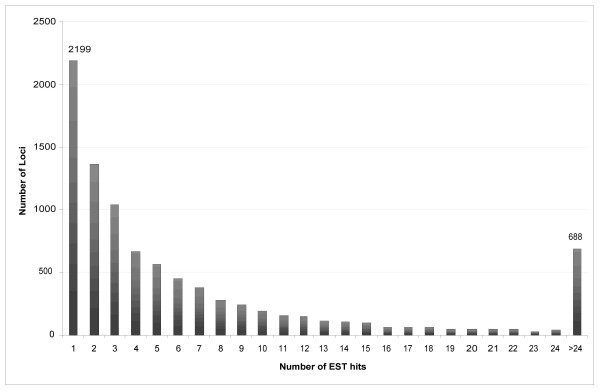
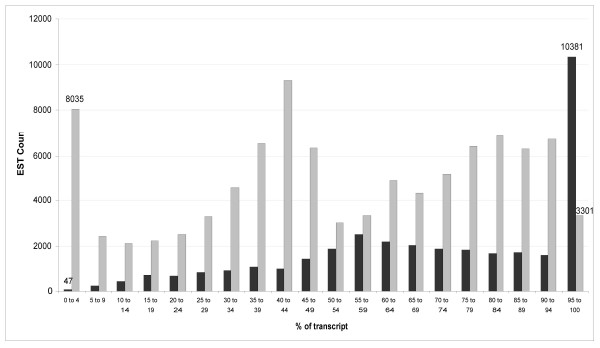
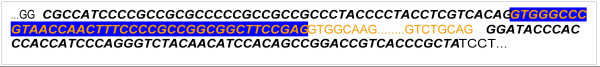
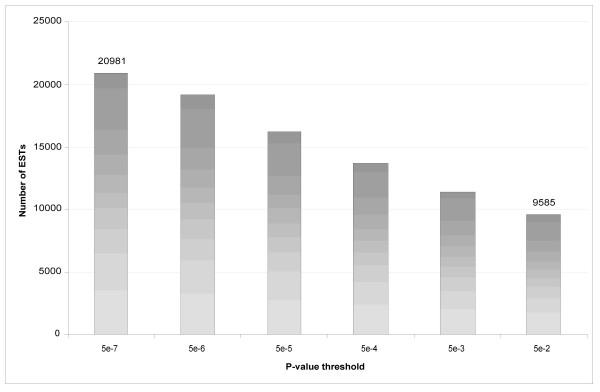
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“Transcript abundance is directly proportional to the number of sequencing reads that map to a specific transcript, resulting in lower falsenegative and false-positive discovery rates in addition to a greater linear range (Bainbridge et al., 2006;Cloonan et al., 2008;Marioni et al., 2008;Mortazavi et al., 2008;Nagalakshmi et al., 2008;Sultan et al., 2008;Wilhelm et al., 2008;Trapnell et al., 2010;W”
From An RNA-Sequencing Transcriptome and Splicing Database of Glia, Neurons, and Vascular Cells of the Cerebral Cortex (2014) · cites this paper as Bainbridge et al., 2006
Experimental — accuracy not yet measured
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Gene Expression Omnibus: NCBI gene expression and hybridization array data repository
Edgar, R. (2002) · Nucleic Acids Research · doi:10.1093/nar/30.1.207 · Cited by 13,288
<tt>BLAT</tt> —The <tt>BLAST</tt> -Like Alignment Tool
Kent, W. J. (2002) · Genome Research · doi:10.1101/gr.229202 · Cited by 8,480
Genome sequencing in microfabricated high-density picolitre reactors
Margulies, M., Egholm, M., Altman, W. E., Attiya, S., Bader, J. S., Bemben, L. A., Berka, J., Braverman, M. S., Chen, Y.-J., Chen, Z., et al. (2005) · Nature · doi:10.1038/nature03959 · Cited by 7,693
Base-Calling of Automated Sequencer Traces Using Phred. I. Accuracy AssessmentPreprint
Ewing, B., Hillier, L., Wendl, M. C., & Green, P. (1998) · Genome Research · doi:10.1101/gr.8.3.175 · Cited by 6,993
Base-Calling of Automated Sequencer Traces Using Phred. II. Error Probabilities
Ewing, B., & Green, P. (1998) · Genome Research · doi:10.1101/gr.8.3.186 · Cited by 5,439
Serial Analysis of Gene Expression
Velculescu, V. E., Zhang, L., Vogelstein, B., & Kinzler, K. W. (1995) · Science · doi:10.1126/science.270.5235.484 · Cited by 3,713
InterProScan: protein domains identifier
Quévillon, E., Silventoinen, V., Pillai, S., Harte, N., Mulder, N., Apweiler, R., & López, R. (2005) · Nucleic Acids Research · doi:10.1093/nar/gki442 · Cited by 2,976
Expression monitoring by hybridization to high-density oligonucleotide arrays
Lockhart, D. J., Dong, H., Byrne, M. C., Follettie, M. T., Gallo, M. V., Chee, M. S., Mittmann, M., Wang, C., Kobayashi, M., Norton, H., et al. (1996) · Nature Biotechnology · doi:10.1038/nbt1296-1675 · Cited by 2,841
Gene expression analysis by massively parallel signature sequencing (MPSS) on microbead arrays.
Brenner, S., Johnson, M., Bridgham, J., Golda, G., Lloyd, D. H., Johnson, D., Luo, S., McCurdy, S., Foy, M., Ewan, M., et al. (2000) · Nature Biotechnology · doi:10.1038/76469 · Cited by 1,360
dbEST — database for “expressed sequence tags”
Boguski, M. S., Lowe, T. M., & Tolstoshev, C. M. (1993) · Nature Genetics · doi:10.1038/ng0893-332 · Cited by 1,340
Research works printed but not linked: 1
- dbEST: [http://www.ncbi.nlm.nih.gov/dbEST/].
Cited by
Sorted by citations
Zhang, Y., Chen, K., Sloan, S. A., Bennett, M. L., Scholze, A. R., O'Keeffe, S., Phatnani, H. P., Guarnieri, P., Caneda, C., Ruderisch, N., et al. (2014) · The Journal of Neuroscience · doi:10.1523/jneurosci.1860-14.2014 · Cited by 4,875
Next-generation DNA sequencing
Shendure, J., & Ji, H. (2008) · Nature Biotechnology · doi:10.1038/nbt1486 · Cited by 4,509
Pan, Q., Shai, O., Lee, L. J., Frey, B. J., & Blencowe, B. J. (2008) · Nature Genetics · doi:10.1038/ng.259 · Cited by 3,951
Next-Generation DNA Sequencing Methods
Mardis, E. R. (2008) · Annual Review of Genomics and Human Genetics · doi:10.1146/annurev.genom.9.081307.164359 · Cited by 2,469
Self-citationApplications of next-generation sequencing technologies in functional genomics
Morozova, O., & Marra, M. A. (2008) · Genomics · doi:10.1016/j.ygeno.2008.07.001 · Cited by 1,222
Self-citation
Metadata
- Authors
- Bainbridge, M. N., Warren, R. L., Hirst, M., Romanuik, T., Zeng, T., Go, A., Delaney, A., Griffith, M., Hickenbotham, M., Magrini, V., Mardis, E. R., Sadar, M. D., Siddiqui, A. S., Marra, M. A., & Jones, S. J.
- Affiliations
- BC Cancer Agency (CA)
- Washington University in St. Louis (US)
- Corresponding author
- Jones, Steven JM
- ORCID
- Venue
- BMC Genomics
- Published
- 2006-09-29
- Volume
- 7
- Issue
- 1
- Pages
- 246-246
- ISSN
- 1471-2164 (ISSN-L)
- PMC
- PMC1592491
- OpenAlex
- W2125013165
- Cited by
- 222
- References
- 33
- Citations by year
- 2026: 6 · 2025: 6 · 2024: 15 · 2023: 15 · 2022: 12 · 2021: 21 · 2020: 8 · 2019: 10 · 2018: 6 · 2017: 6 · 2016: 3 · 2015: 10 · 2014: 5 · 2013: 9 · 2012: 8
- FWCI
- 7.62
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- Locations
- BMC Genomics (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed (publishedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- Funders
- Michael Smith Health Research BC
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene expression and cancer classification
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Keywords
- LNCaP cells, prostate cancer, gene expression profiling, sequencing by synthesis, expressed sequence tags, alternative splicing, single nucleotide polymorphisms, R1881, androgen therapy, novel transcripts, genome mapping
- MeSH terms
- Adenocarcinoma, Androgens, Chromosome Mapping, Chromosomes, Human, Exons, Humans, Male, Neoplasms, Hormone-Dependent, Prostatic Neoplasms, RNA, Messenger, RNA, Neoplasm, Sequence Homology, Nucleic Acid, Transcription, Genetic, Metribolone, Gene Expression Regulation, Neoplastic, Sequence Alignment, Alternative Splicing, Sequence Analysis, DNA, DNA, Complementary, Expressed Sequence Tags, Polymorphism, Single Nucleotide, Gene Expression Profiling, Cell Line, Tumor
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
