

The DOI matches its Crossref record; 10 of the 10 references it cites were found in a registry or an index.
Tools and best practices for data processing in allelic expression analysis.
Open accessPublished version of a preprint
Abstract
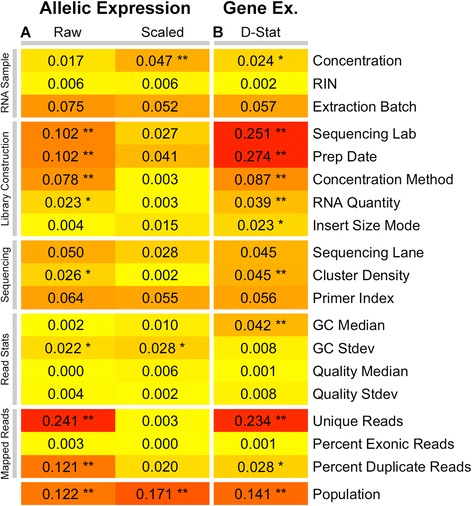
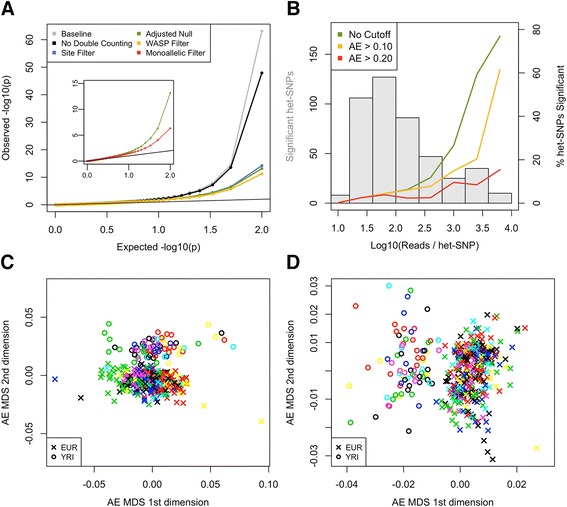
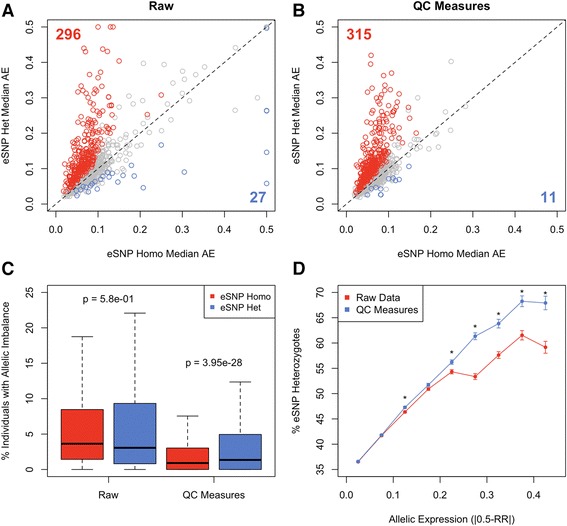
ABSTRACT Allelic expression (AE) analysis has become an important tool for integrating genome and transcriptome data to characterize various biological phenomena such as cis -regulatory variation and nonsense-mediated decay. In this paper, we systematically analyze the properties of AE read count data and technical sources of error, such as low-quality or double-counted RNA-seq reads, genotyping errors, allelic mapping bias, and technical covariates due to sample preparation and sequencing, and variation in total read depth. We provide guidelines for correcting and filtering for such errors, and show that the resulting AE data has extremely low technical noise. Finally, we introduce novel software for high-throughput production of AE data from RNA-sequencing data, implemented in the GATK framework. These improved tools and best practices for AE analysis yield higher quality AE data by reducing technical bias. This provides a practical framework for wider adoption of AE analysis by the genomics community.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“The quality of the genotype calls is crucial for allele assignment, so the genotype data needs to be collected carefully and quality control and filtering is required to avoid biases and false positive hits 9.”
From SNPsplit: Allele-specific splitting of alignments between genomes with known SNP genotypes (2016) · cites this paper as [9]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Van der Auwera, G. A., Carneiro, M. O., Hartl, C., Poplin, R., del Angel, G., Levy‐Moonshine, A., Jordan, T., Shakir, K., Roazen, D., Thibault, J., et al. (2013) · Current Protocols in Bioinformatics · doi:10.1002/0471250953.bi1110s43 · Cited by 6,687
Self-citationThe Genotype-Tissue Expression (GTEx) pilot analysis: Multitissue gene regulation in humans
The GTEx Consortium, Ardlie, K. G., Deluca, D. S., Segrè, A. V., Sullivan, T. J., Young, T. R., Gelfand, E. T., Trowbridge, C. A., Maller, J. B., Tukiainen, T., et al. (2015) · Science · doi:10.1126/science.1262110 · Cited by 5,062
Self-citationTranscriptome and genome sequencing uncovers functional variation in humans
The Geuvadis Consortium, Lappalainen, T., Sammeth, M., Friedländer, M. R., ‘t Hoen, P. A. C., Monlong, J., Rivas, M. A., Gonzàlez-Porta, M., Kurbatova, N., Griebel, T., et al. (2013) · Nature · doi:10.1038/nature12531 · Cited by 1,888
Self-citationUnderstanding mechanisms underlying human gene expression variation with RNA sequencing
Pickrell, J. K., Marioni, J. C., Pai, A. A., Degner, J. F., Engelhardt, B. E., Nkadori, E., Veyrieras, J.-B., Stephens, M., Gilad, Y., & Pritchard, J. K. (2010) · Nature · doi:10.1038/nature08872 · Cited by 1,256
Transcriptome genetics using second generation sequencing in a Caucasian population
Montgomery, S. B., Sammeth, M., Gutierrez-Arcelus, M., Lach, R. P., Ingle, C., Nisbett, J., Guigo, R., & Dermitzakis, E. T. (2010) · Nature · doi:10.1038/nature08903 · Cited by 797
Effect of read-mapping biases on detecting allele-specific expression from RNA-sequencing data.
Degner, J. F., Marioni, J. C., Pai, A. A., Pickrell, J. K., Nkadori, E., Gilad, Y., & Pritchard, J. K. (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp579 · Cited by 502
Passive and active DNA methylation and the interplay with genetic variation in gene regulation
Gutierrez-Arcelus, M., Lappalainen, T., Montgomery, S. B., Buil, A., Ongen, H., Yurovsky, A., Bryois, J., Giger, T., Romano, L., Planchon, A., et al. (2013) · eLife · doi:10.7554/elife.00523 · Cited by 440
Self-citationEffect of predicted protein-truncating genetic variants on the human transcriptome
Rivas, M. A., Pirinen, M., Conrad, D. F., Lek, M., Tsang, E. K., Karczewski, K. J., Maller, J. B., Kukurba, K. R., DeLuca, D. S., Fromer, M., et al. (2015) · Science · doi:10.1126/science.1261877 · Cited by 313
Self-citationReproducibility of high-throughput mRNA and small RNA sequencing across laboratories
The GEUVADIS Consortium, 't Hoen, P. A. C., Friedländer, M. R., Almlöf, J., Sammeth, M., Pulyakhina, I., Anvar, S. Y., Laros, J. F. J., Buermans, H. P. J., Karlberg, O., et al. (2013) · Nature Biotechnology · doi:10.1038/nbt.2702 · Cited by 250
Self-citationStevenson, K. R., Coolon, J. D., & Wittkopp, P. J. (2013) · BMC Genomics · doi:10.1186/1471-2164-14-536 · Cited by 140
Research works printed but not linked: 32
- Adoue V, Schiavi A, Light N, Almlöf JC, Lundmark P, Ge B, et al. Allelic expression mapping across cellular lineages to establish impact of non-coding SNPs. Mol Syst Biol. 2014;10:754. · DOI
- Buil A, Brown AA, Lappalainen T, Viñuela A, Davies MN, Zheng H-F, et al. Gene-gene and gene-environment interactions detected by transcriptome sequence analysis in twins. Nat Genet. 2015;47:88–91. · DOI
- Ge B, Pokholok DK, Kwan T, Grundberg E, Morcos L, Verlaan DJ, et al. Global patterns of cis variation in human cells revealed by high-density allelic expression analysis. Nat Genet. 2009;41:1216–22. · DOI
- Battle A, Mostafavi S, Zhu X, Potash JB, Weissman MM, McCormick C, et al. Characterizing the genetic basis of transcriptome diversity through RNA-sequencing of 922 individuals. Genome Res. 2014;24:14–24. · DOI · Paper page
- Kukurba KR, Zhang R, Li X, Smith KS, Knowles DA, How Tan M, et al. Allelic expression of deleterious protein-coding variants across human tissues. PLoS Genet. 2014;10, e1004304. · DOI
- MacArthur DG, Balasubramanian S, Frankish A, Huang N, Morris J, Walter K, et al. A systematic survey of loss-of-function variants in human protein-coding genes. Science. 2012;335:823–8. · DOI · Paper page
- Montgomery SB, Lappalainen T, Gutierrez-Arcelus M, Dermitzakis ET. Rare and common regulatory variation in population-scale sequenced human genomes. PLoS Genet. 2011;7, e1002144. · DOI
- Baran Y, Subramaniam M, Biton A, Tukiainen T, Tsang EK, Rivas MA, et al. The landscape of genomic imprinting across diverse adult human tissues. Genome Res. 2015;25:927–36. · DOI
- Morcos L, Ge B, Koka V, Lam KCL, Pokholok DK, Gunderson KL, et al. Genome-wide assessment of imprinted expression in human cells. Genome Biol. 2011;12:R25. · DOI
- Gimelbrant A, Hutchinson JN, Thompson BR, Chess A. Widespread monoallelic expression on human autosomes. Science. 2007;318:1136–40. · DOI · Paper page
- Borel C, Ferreira PG, Santoni F, Delaneau O, Fort A, Popadin KY, et al. Biased allelic expression in human primary fibroblast single cells. Am J Hum Genet. 2015;96:70–80. · DOI · Paper page
- Soderlund CA, Nelson WM, Goff SA. Allele Workbench: transcriptome pipeline and interactive graphics for allele-specific expression. PLoS One. 2014;9, e115740. · DOI
- Leung D, Jung I, Rajagopal N, Schmitt A, Selvaraj S, Lee AY, et al. Integrative analysis of haplotype-resolved epigenomes across human tissues. Nature. 2015;518:350–4. · DOI
- Li H, Su X, Gallegos J, Lu Y, Ji Y, Molldrem JJ, et al. dsPIG: a tool to predict imprinted genes from the deep sequencing of whole transcriptomes. BMC Bioinformatics. 2012;13:271. · DOI
- Pirinen M, Lappalainen T, Zaitlen NA, GTEx Consortium, Dermitzakis ET, Donnelly P, et al. Assessing allele-specific expression across multiple tissues from RNA-seq read data. Bioinformatics. 2015;31:2497–504. · DOI
- Cho H, Davis J, Li X, Smith KS, Battle A, Montgomery SB. High-resolution transcriptome analysis with long-read RNA sequencing. PLoS One. 2014;9, e108095. · DOI
- Turro E, Su S-Y, Gonçalves Â, Coin LJM, Richardson S, Lewin A. Haplotype and isoform specific expression estimation using multi-mapping RNA-seq reads. Genome Biol. 2011;12:R13. · DOI · Paper page
- McKenna A, Hanna M, Banks E, Sivachenko A, Cibulskis K, Kernytsky A, et al. The Genome Analysis Toolkit: a MapReduce framework for analyzing next-generation DNA sequencing data. Genome Res. 2010;20:1297–303. · DOI · Paper page
- DePristo MA, Banks E, Poplin R, Garimella KV, Maguire JR, Hartl C, et al. A framework for variation discovery and genotyping using next-generation DNA sequencing data. Nat Genet. 2011;43:491–8. · DOI
- Romanel A, Lago S, Prandi D, Sboner A, Demichelis F. ASEQ: fast allele-specific studies from next-generation sequencing data. BMC Med Genomics. 2015;8:9. · DOI
- Panousis NI, Gutierrez-Arcelus M, Dermitzakis ET, Lappalainen T. Allelic mapping bias in RNA-sequencing is not a major confounder in eQTL studies. Genome Biol. 2014;15:467. · DOI
- Gutierrez-Arcelus M, Ongen H, Lappalainen T, Montgomery SB, Buil A, Yurovsky A, et al. Tissue-specific effects of genetic and epigenetic variation on gene regulation and splicing. PLoS Genet. 2015;11, e1004958. · DOI
- Kilpinen H, Waszak SM, Gschwind AR, Raghav SK, Witwicki RM, Orioli A, et al. Coordinated effects of sequence variation on DNA binding, chromatin structure, and transcription. Science. 2013;342:744–7. · DOI · Paper page
- Gerstein MB, Kundaje A, Hariharan M, Landt SG, Yan K-K, Cheng C, et al. Architecture of the human regulatory network derived from ENCODE data. Nature. 2012;489:91–100. · DOI
- Rozowsky J, Abyzov A, Wang J, Alves P, Raha D, Harmanci A, et al. AlleleSeq: analysis of allele-specific expression and binding in a network framework. Mol Syst Biol. 2011;7:522–2. · DOI · Paper page
- Wu TD, Nacu S. Fast and SNP-tolerant detection of complex variants and splicing in short reads. Bioinformatics. 2010;26:873–81. · DOI · Paper page
- van de Geijn B, McVicker G, Gilad Y, Pritchard J. WASP: allele-specific software for robust discovery of molecular quantitative trait loci. bioRxiv. 2014. http://dx.doi.org/10.1101/011221 . · DOI
- Piskol R, Ramaswami G, Li JB. Reliable identification of genomic variants from RNA-seq data. Am J Hum Genet. 2013;93:641–51. · DOI
- Deelen P, Zhernakova DV, de Haan M, van der Sijde M, Bonder MJ, Karjalainen J, et al. Calling genotypes from public RNA-sequencing data enables identification of genetic variants that affect gene-expression levels. Genome Med. 2015;7:30. · DOI
- Kumasaka N, Knights A, Gaffney D. Fine-mapping cellular QTLs with RASQUAL and ATAC-seq. bioRxiv. 2015. http://dx.doi.org/10.1101/018788 . · DOI
- Skelly DA, Johansson M, Madeoy J, Wakefield J, Akey JM. A powerful and flexible statistical framework for testing hypotheses of allele-specific gene expression from RNA-seq data. Genome Res. 2011;21:1728–37. · DOI · Paper page
- Zhang R, Li X, Ramaswami G, Smith KS, Turecki G, Montgomery SB, et al. Quantifying RNA allelic ratios by microfluidic multiplex PCR and sequencing. Nat Methods. 2014;11:51–4. · DOI
Cited by
Sorted by citations
The GTEx Consortium atlas of genetic regulatory effects across human tissues
The GTEx Consortium, Aguet, F., Anand, S., Ardlie, K. G., Gabriel, S., Getz, G. A., Graubert, A., Hadley, K., Handsaker, R. E., Huang, K. H., et al. (2020) · Science · doi:10.1126/science.aaz1776 · Cited by 5,547
Self-citationGenetic effects on gene expression across human tissues
GTEx Consortium (2017) · Nature · doi:10.1038/nature24277 · Cited by 4,036
“1 ×10⁻³ for (75,100]% vs [0,25]%, (75,100 vs (25,50]% and (75,100]% vs (50,75]%, respectively).”
Landscape of X chromosome inactivation across human tissues
GTEx Consortium, Tukiainen, T., Villani, A.-C., Yen, A., Rivas, M. A., Marshall, J. L., Satija, R., Aguirre, M., Gauthier, L., Fleharty, M., et al. (2017) · Nature · doi:10.1038/nature24265 · Cited by 1,138
“The analysis was restricted to biallelic SNPs due to known issues in mapping bias in RNA-seq against indels 37 .”
InfluentialSelf-citationImproving genetic diagnosis in Mendelian disease with transcriptome sequencing
Cummings, B. B., Marshall, J. L., Tukiainen, T., Lek, M., Donkervoort, S., Foley, A. R., Bolduc, V., Waddell, L. B., Sandaradura, S. A., O’Grady, G. L., et al. (2017) · Science Translational Medicine · doi:10.1126/scitranslmed.aal5209 · Cited by 831
“e (38) d(39)m,(40) asja (41) dda (42) kja (43) jj (44) kl (45) jk (46)”
Translating RNA sequencing into clinical diagnostics: opportunities and challenges
Byron, S. A., Keuren‐Jensen, K. R. V., Engelthaler, D. M., Carpten, J. D., & Craig, D. W. (2016) · Nature Reviews Genetics · doi:10.1038/nrg.2016.10 · Cited by 823
“Recent reports provide recommendations for bioinformatic analysis and data processing to address these and other challenges and introduce tools for improved detection of ASE from RNA-seq data 42, 48 .”
Background
Metadata
- Authors
- Castel, S. E., Levy-Moonshine, A., Mohammadi, P., Banks, E., & Lappalainen, T.
- Affiliations
- New York Genome Center (US)
- Broad Institute (US)
- Corresponding author
- Castel, Stephane E.
- Lappalainen, Tuuli
- Venue
- Genome Biology
- Published
- 2015-09-17
- Volume
- 16
- Issue
- 1
- Pages
- 195-195
- ISSN
- 1465-6906 (ISSN-L)
- 1474-760X
- PMC
- PMC4574606
- OpenAlex
- W2142933606
- Cited by
- 375
- References
- 43
- Citations by year
- 2026: 29 · 2025: 36 · 2024: 25 · 2023: 45 · 2022: 25 · 2021: 47 · 2020: 33 · 2019: 35 · 2018: 37 · 2017: 38 · 2016: 21 · 2015: 4
- FWCI
- 14.48
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- Locations
- Genome biology (publishedVersion, cc-by)
- bioRxiv (Cold Spring Harbor Laboratory) (acceptedVersion, other-oa)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Funders
- University of Miami (MH090936, MH090937, MH090951, MH090948, DA006227, MH090941)
- Harvard University (MH090948)
- University of Chicago (MH090951, MH090937)
- Broad Institute (HHSN268201000029C)
- New York Genome Center
- Université de Genève (MH090941)
- National Institutes of Health (10XS170, #HHSN261200800001E, HHSN268201000029C, 10XS171, HHSN261200800001C, 10ST1035, MH090937)
- National Heart, Lung, and Blood Institute (HHSN268201000029C, HHSN261200800001E)
- National Institute of Mental Health (R01MH101814, R01MH090937, HHSN261200800001E, R01MH090948, R01MH090941, R01MH090936, R01MH090951)
- National Institute on Drug Abuse (DA006227, R01DA006227)
- National Human Genome Research Institute (U01HG006569, HHSN261200800001E)
- NIH Office of the Director (HHSN261200800001E)
- National Cancer Institute (HHSN261200800001E))
- National Institute of Neurological Disorders and Stroke
- Common Fund
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › RNA Research and Splicing
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Keywords
- allelic expression, RNA-seq, quality control, genotyping errors, cis-regulatory variation, nonsense-mediated mRNA decay, bioinformatics software
- MeSH terms
- Alleles, Cell Line, Data Interpretation, Statistical, Humans, Software, Gene Expression, Sequence Analysis, RNA, Gene Expression Profiling, Genotyping Techniques
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
