

The DOI matches its Crossref record; 155 of the 155 references it cites were found in a registry or an index.
Computational methods for chromosome-scale haplotype reconstruction
Open accessAbstract
High-quality chromosome-scale haplotype sequences of diploid genomes, polyploid genomes, and metagenomes provide important insights into genetic variation associated with disease and biodiversity. However, whole-genome short read sequencing does not yield haplotype information spanning whole chromosomes directly. Computational assembly of shorter haplotype fragments is required for haplotype reconstruction, which can be challenging owing to limited fragment lengths and high haplotype and repeat variability across genomes. Recent advancements in long-read and chromosome-scale sequencing technologies, alongside computational innovations, are improving the reconstruction of haplotypes at the level of whole chromosomes. Here, we review recent and discuss methodological progress and perspectives in these areas.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
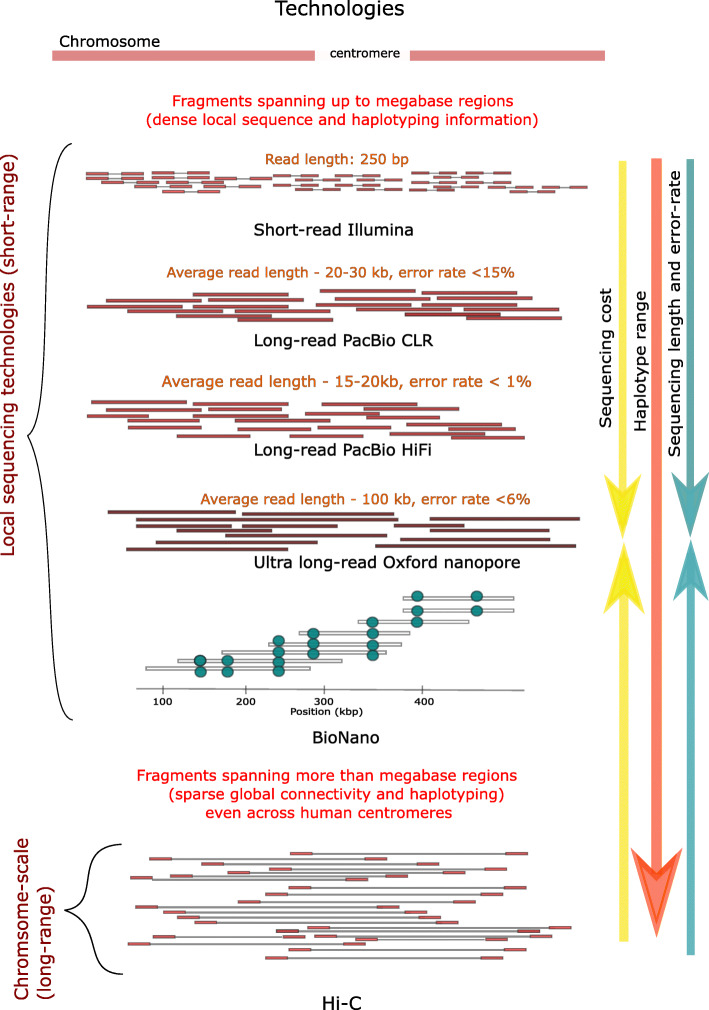
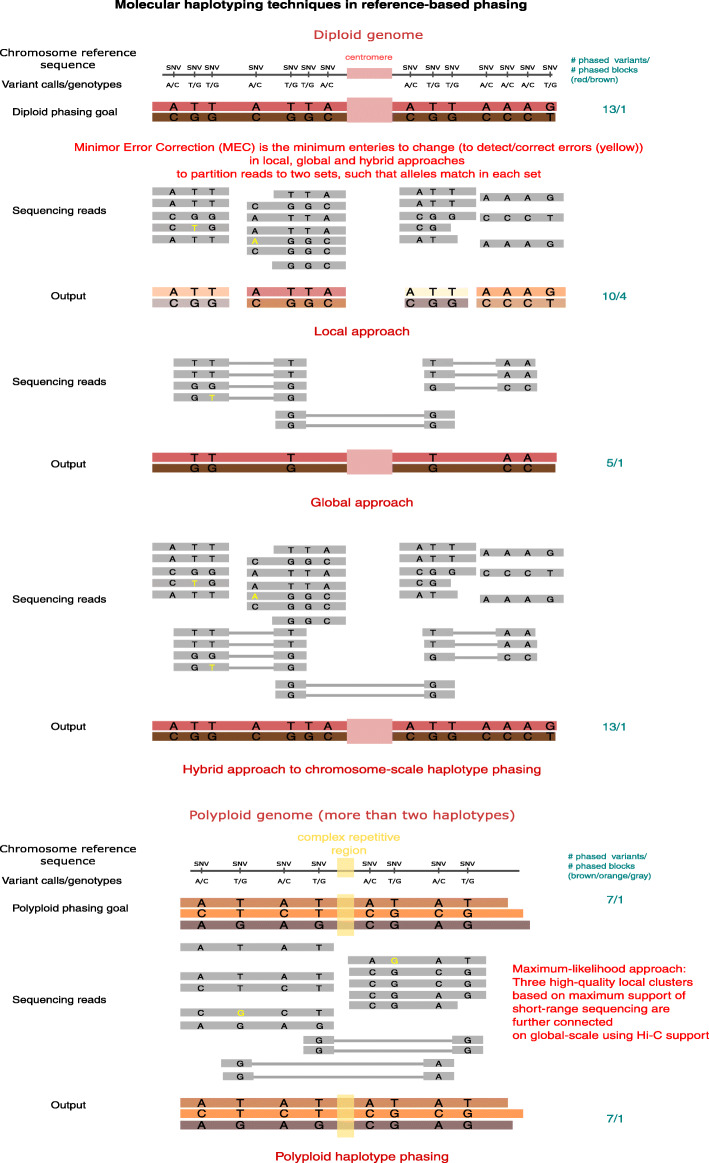
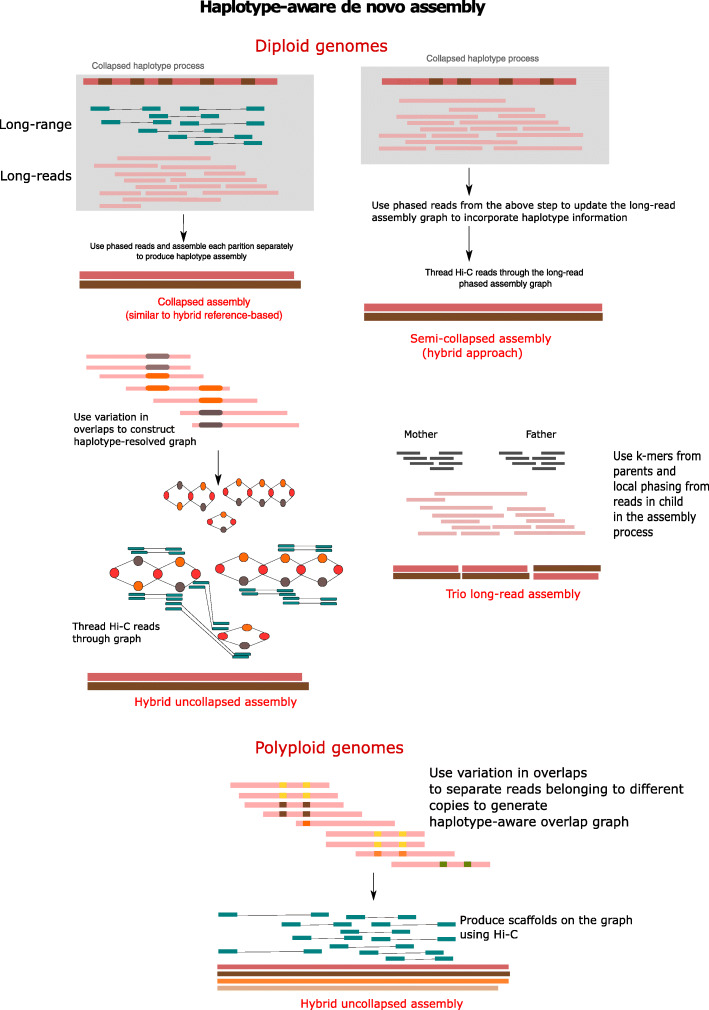
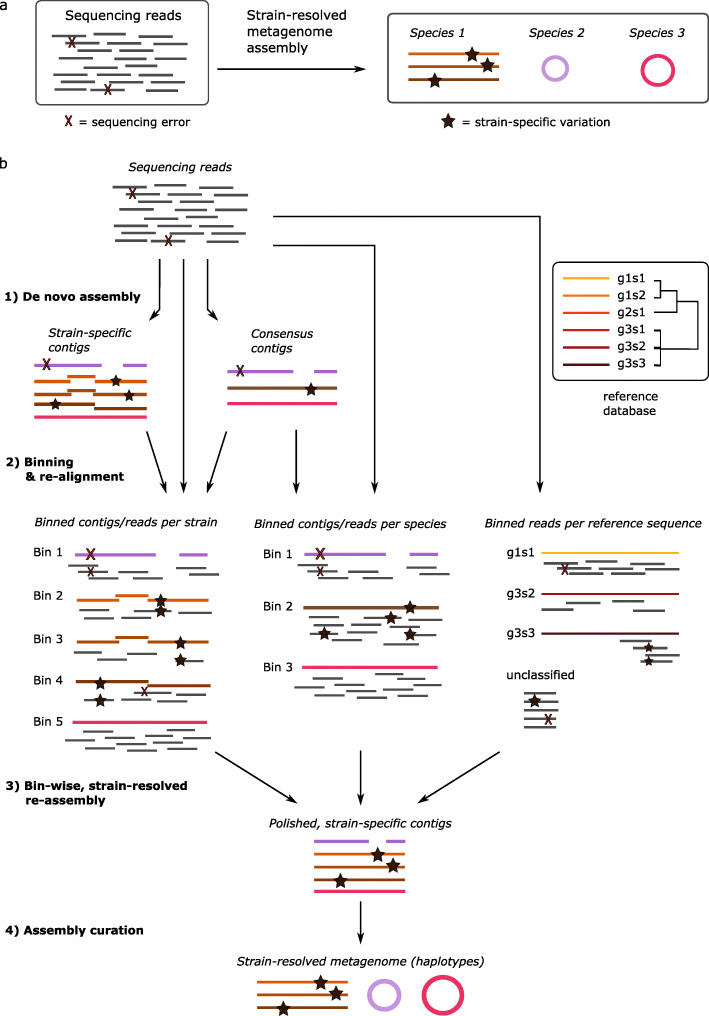
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The check wasn’t confident either way, so it gives no verdict — never disputed.
In the citing paper
“Pstools reported slightly lower switch error rates than Yak but comparable Hamming error rates (Supplementary Fig. 2).”
From A draft human pangenome reference. (2023) · cites this paper as [23]
Experimental — accuracy not yet measured
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Structure, function and diversity of the healthy human microbiome
The Human Microbiome Project Consortium (2012) · Nature · doi:10.1038/nature11234 · Cited by 11,726
Li, D., Liu, C.-M., Luo, R., Sadakane, K., & Lam, T. (2015) · Bioinformatics · doi:10.1093/bioinformatics/btv033 · Cited by 10,133
Santos, A. G., da Rocha, G. O., & de Andrade, J. B. (2019) · Scientific Reports · doi:10.1038/s41598-018-37186-2 · Cited by 8,495
Canu: scalable and accurate long-read assembly via adaptive k -mer weighting and repeat separationPreprint
Koren, S., Walenz, B. P., Berlin, K., Miller, J. R., Bergman, N. H., & Phillippy, A. M. (2017) · Genome Research · doi:10.1101/gr.215087.116 · Cited by 7,260
Assembly of long, error-prone reads using repeat graphs
Kolmogorov, M., Yuan, J., Lin, Y., & Pevzner, P. A. (2019) · Nature Biotechnology · doi:10.1038/s41587-019-0072-8 · Cited by 5,986
metaSPAdes: a new versatile metagenomic assembler
Nurk, S., Meleshko, D., Korobeynikov, A., & Pevzner, P. A. (2017) · Genome Research · doi:10.1101/gr.213959.116 · Cited by 4,496
Mash: fast genome and metagenome distance estimation using MinHash
Ondov, B. D., Treangen, T. J., Melsted, P., Mallonee, A. B., Bergman, N. H., Koren, S., & Phillippy, A. M. (2016) · Genome biology · doi:10.1186/s13059-016-0997-x · Cited by 3,373
Browning, S. R., & Browning, B. L. (2007) · The American Journal of Human Genetics · doi:10.1086/521987 · Cited by 3,217
Fast and accurate de novo genome assembly from long uncorrected readsPreprint
Vaser, R., Sović, I., Nagarajan, N., & Šikić, M. (2017) · Genome Research · doi:10.1101/gr.214270.116 · Cited by 3,059
Peng, Y., Leung, H. C. M., Yiu, S., & Chin, F. Y. L. (2012) · Bioinformatics · doi:10.1093/bioinformatics/bts174 · Cited by 2,998
Research works printed but not linked: 3
- Zaccaria S, Raphael BJ. Characterizing the allele- and haplotype-specific copy number landscape of cancer genomes at single-cell resolution with CHISEL. bioRxiv. 2019:837195. https://doi.org/10.1101/837195. · DOI
- Kronenberg ZN, et al. FALCON-Phase: Integrating PacBio and Hi-C data for phased diploid genomes. bioRxiv. 2018:327064. https://doi.org/10.1101/327064. · DOI
- Bzikadze, A. V. & Pevzner, P. A. centroFlye: assembling centromeres with long error-prone reads. 2019; 772103 doi:https://doi.org/10.1101/772103. · DOI
Cited by
Sorted by citations
Items 101–116 of 116
Chen, H., Nie, F., Luo, F., & Wang, J. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.11.07.687303
Zumaila, F., Jeevalatha, A., & Biju, C. N. (2025) · Fungal Biology · doi:10.1016/j.funbio.2025.101693
Luo, X., Kang, X., & Schönhuth, A. (2022) · Figshare · doi:10.6084/m9.figshare.18849306
Luo, X., Kang, X., & Schönhuth, A. (2022) · Figshare · doi:10.6084/m9.figshare.18849300
Additional file 1 of GreenHill: a de novo chromosome-level scaffolding and phasing tool using Hi-C
Ouchi, S., Kajitani, R., & Itoh, T. (2023) · Figshare · doi:10.6084/m9.figshare.23665093
Additional file 2 of GreenHill: a de novo chromosome-level scaffolding and phasing tool using Hi-C
Ouchi, S., Kajitani, R., & Itoh, T. (2023) · Figshare · doi:10.6084/m9.figshare.23665096
Huang, B., Wei, G., Wang, B., Ju, F., Zhong, Y., Shi, Z., Sun, S., & Bu, D. (2021) · Figshare · doi:10.6084/m9.figshare.16910035
An accurate phasing tool via high-quality heterozygous variationsPreprint
Zhang, J., Nie, F., & Wang, J. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.64898/2025.12.21.695866
“The nine datasets were aligned to the human reference genome (GRCh38) using minimap2 (Li, 2018, 2021), with parameters optimized for PacBio HiFi and Nanopore reads.”
nTChap: an accurate method for polyploid haplotype reconstruction
Gao, Y., Qi, J., Yu, T., & Li, G. (2026) · BMC Genomics · doi:10.1186/s12864-025-12440-w
“This approach lays the first theoretical foundation of polyploidy phasing problems [6].”
Does Modern Plant Breeding Techniques Increase Crop Stability Performance?
Ćeran, M., Đorđević, V., Miladinović, J., Miladinović, D., Trkulja, D., Radanović, A., & Kondić‐Špika, A. (2026) · Springer eBooks · doi:10.1007/978-981-95-5664-9_5
Lv, W., Zou, Y., Shang, Z., Wei, S., Sun, C., Xu, J., Chen, H., Tao, J., Tian, H., Bi, S., et al. (2026) · Genome biology · doi:10.1186/s13059-026-04019-9
“The concepts of haplotypes and blocks are formed when the alleles of more than two variant sites exhibit a higher degree of LD compared to other regions of the genome which is crucial to better understanding of human LD patterns [3–5].”
Lei, F., Li, D., Zhu, H., Xia, Y., Chen, X. D., Yu, J., Guo, S., & Yu, J. Q. (2026) · Evolutionary Applications · doi:10.1111/eva.70229
“Haplotypes can retain the integrity of a segment of genetic information and can be used to represent a specific type (Garg 2021; Zhang et al. 2024).”
Magnitov, M. D., van der Weide, R. H., Witvliet, A. F., Hernández-Quiles, M., Martinovic, M., Teunissen, H., Braccioli, L., Vermeulen, M., & de Wit, E. (2026) · Nature Communications · doi:10.1038/s41467-026-72392-x
“Previous studies have shown that a combination of dense local and sparse chromosome-scale phasing methods produces the highest-quality haplotypes 60 .”
Piet, Q., Besse, P., Bocs, S., Bouchez, O., Bourge, M., Charron, C., Cravero, C., Dedours, A., Droc, G., Dron, M., et al. (2026) · BMC Genomics · doi:10.1186/s12864-026-12926-1
“The discrepancy between the CR0040 V1 and Daphna assembly lengths was attributed to the impact of PE [16], inducing highly unbalanced DNA content within a given tissue and challenging the assembly of sequencing reads with highly unbalanced depths, particularly in low depth repetitive regions [63].”
Antysheva, Z., Bogdanov, V. P., Rutkovskaya, E. A., Stepanova, A., Esibov, A. A., Krupinova, J., Shchekina, V. S., Avsievich, E., Tatyana, F., Bazarova, E., et al. (2026) · International Journal of Molecular Sciences · doi:10.3390/ijms27156938
“Like LRS, SRS allows for read-based phasing; however, as expected, phased blocks are on average shorter [27] and it is unclear if standalone read-based phasing in targeted SRS can be utilized for reliable CAH diagnosis.”
Superior haplotypes for transforming crop breeding
Kumar, N. V., Bogati, A., Singh, U. M., Rajesh, A. P., Srinivas, T., Kohli, A., Kalia, S., Kumar, A., Sinha, P., & Singh, V. K. (2026) · Theoretical and Applied Genetics · doi:10.1007/s00122-026-05337-z
“Tools like BEAGLE use population reference panels for statistical phasing, but only handle short blocks and cannot reliably phase entire chromosomes in polyploid genomes (Garg 2021).”
Metadata
- Authors
- Garg, S.
- Affiliations
- University of Copenhagen (DK)
- Corresponding author
- Garg, Shilpa
- ORCID
- Garg, Shilpa
- Venue
- Genome Biology
- Published
- 2021-04-12
- Volume
- 22
- Issue
- 1
- Pages
- 101-101
- ISSN
- 1465-6906 (ISSN-L)
- 1474-760X
- PMC
- PMC8040228
- OpenAlex
- W3156261638
- Cited by
- 116
- References
- 154
- Citations by year
- 2026: 11 · 2025: 26 · 2024: 25 · 2023: 20 · 2022: 28 · 2021: 6
- FWCI
- 2.07
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Locations
- Genome biology (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Research at the University of Copenhagen (University of Copenhagen) (publishedVersion, cc-by)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Chromosomal and Genetic Variations
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genomic variations and chromosomal abnormalities
- Keywords
- haplotype reconstruction, haplotype assembly, diploid genome, polyploid genomes, metagenome, long-read sequencing, repetitive regions, genetic variation
- MeSH terms
- Chromosomes, Diploidy, Haplotypes, Humans, Polyploidy, Sequence Analysis, DNA, Computational Biology, Genomics, Metagenome, Metagenomics, High-Throughput Nucleotide Sequencing
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
