

The DOI matches its Crossref record; 35 of the 36 references it cites were found in a registry or an index.
ANGSD: Analysis of Next Generation Sequencing Data
Open accessAbstract
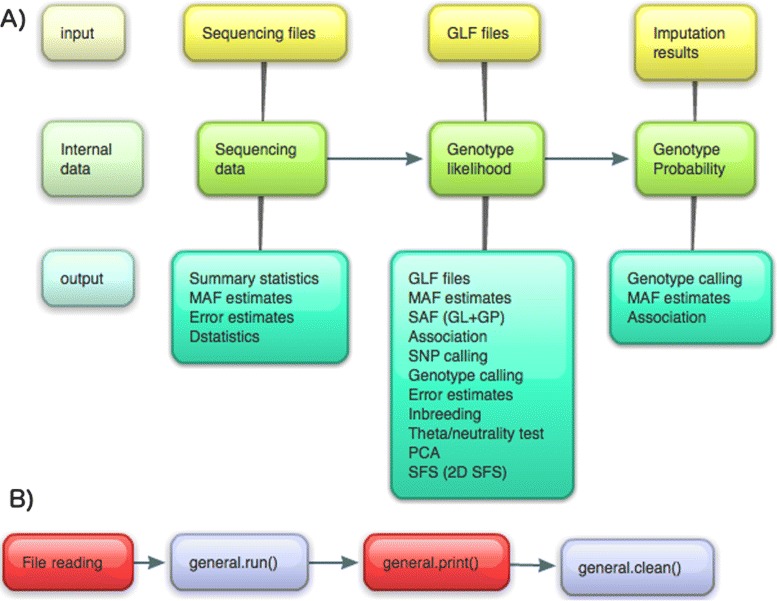
BACKGROUND: High-throughput DNA sequencing technologies are generating vast amounts of data. Fast, flexible and memory efficient implementations are needed in order to facilitate analyses of thousands of samples simultaneously. RESULTS: We present a multithreaded program suite called ANGSD. This program can calculate various summary statistics, and perform association mapping and population genetic analyses utilizing the full information in next generation sequencing data by working directly on the raw sequencing data or by using genotype likelihoods. CONCLUSIONS: The open source c/c++ program ANGSD is available at http://www.popgen.dk/angsd . The program is tested and validated on GNU/Linux systems. The program facilitates multiple input formats including BAM and imputed beagle genotype probability files. The program allow the user to choose between combinations of existing methods and can perform analysis that is not implemented elsewhere.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“we excluded libraries with X-contamination estimates greater than 1.5%32.”
From Ancient DNA from Chalcolithic Israel reveals the role of population mixture in cultural transformation (2018) · cites this paper as [32]
The check wasn’t confident either way, so it gives no verdict — never disputed.
In the citing paper
“We evaluated each individual for evidence of authentic ancient DNA by limiting to libraries with a minimum of 3% cytosine-to-thymine errors at the final nucleotide29, by requiring that the ratio of X-to-Y-chromosome sequences was characteristic of either a male or a female, by requiring >95% matching to the consensus sequence of mitochondrial DNA30, and by requiring (for males) a lack of variation”
From Ancient DNA from Chalcolithic Israel reveals the role of population mixture in cultural transformation (2018) · cites this paper as [32]
Experimental — accuracy not yet measuredThe check wasn’t confident either way, so it gives no verdict — never disputed.
In the citing paper
“We assessed the quality of each library at the screening stage using three standard methods for determining ancient DNA authenticity.”
From Ancient DNA from Chalcolithic Israel reveals the role of population mixture in cultural transformation (2018) · cites this paper as [32]
Experimental — accuracy not yet measuredIn the citing paper
“We next used ANGSD (v0.910) (41), which compares mismatch rates between polymorphic sites on the X chromosome to estimate nuclear contamination in males.”
From Ancient DNA sheds light on the genetic origins of early Iron Age Philistines (2019) · cites this paper as [41]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Fast and accurate short read alignment with Burrows-Wheeler transform
Li, H., & Durbin, R. (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp324 · Cited by 58,607
The Genome Analysis Toolkit: A MapReduce framework for analyzing next-generation DNA sequencing data
McKenna, A., Hanna, M., Banks, E., Sivachenko, A., Cibulskis, K., Kernytsky, A., Garimella, K., Altshuler, D., Gabriel, S., Daly, M., et al. (2010) · Genome Research · doi:10.1101/gr.107524.110 · Cited by 27,895
Ultrafast and memory-efficient alignment of short DNA sequences to the human genome.
Langmead, B., Trapnell, C., Pop, M., & Salzberg, S. L. (2009) · Genome Biology · doi:10.1186/gb-2009-10-3-r25 · Cited by 21,849
A map of human genome variation from population-scale sequencing.
The 1000 Genomes Project Consortium (2010) · Nature · doi:10.1038/nature09534 · Cited by 7,877
Li, H. (2011) · Bioinformatics · doi:10.1093/bioinformatics/btr509 · Cited by 6,454
A second generation human haplotype map of over 3.1 million SNPs
The International HapMap Consortium (2007) · Nature · doi:10.1038/nature06258 · Cited by 4,438
A draft sequence of the Neandertal genome
Green, R. E., Krause, J., Briggs, A. W., Maricic, T., Stenzel, U., Kircher, M., Patterson, N., Li, H., Zhai, W., Fritz, M. H.-Y., et al. (2010) · Science · doi:10.1126/science.1188021 · Cited by 4,318
Self-citationSOAP2: an improved ultrafast tool for short read alignment
Li, R., Yu, C., Li, Y., Lam, T., Yiu, S., Kristiansen, K., & Wang, J. (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp336 · Cited by 3,763
Ancient Admixture in Human History
Patterson, N., Moorjani, P., Luo, Y., Mallick, S., Rohland, N., Zhan, Y., Genschoreck, T., Webster, T., & Reich, D. (2012) · Genetics · doi:10.1534/genetics.112.145037 · Cited by 2,848
Genetic history of an archaic hominin group from Denisova Cave in Siberia
Reich, D., Green, R. E., Kircher, M., Krause, J., Patterson, N., Durand, E. Y., Viola, B., Briggs, A. W., Stenzel, U., Johnson, P. L. F., et al. (2010) · Nature · doi:10.1038/nature09710 · Cited by 1,991
Research works printed but not linked: 1
- Li H, Durbin R: Fast and accurate short read alignment with Burrows-Wheeler transform . Bioinformatics. 2009, 25: 1754-1760. 10.1093/bioinformatics/btp324. · DOI · Paper page
Cited by
Sorted by citations
Population genomics of Bronze Age Eurasia
Allentoft, M. E., Sikora, M., Sjögren, K.-G., Rasmussen, S., Rasmussen, M., Stenderup, J., Damgaard, P. B., Schroeder, H., Ahlström, T., Vinner, L., et al. (2015) · Nature · doi:10.1038/nature14507 · Cited by 1,550
Self-citationGenome-wide patterns of selection in 230 ancient Eurasians
Mathieson, I., Lazaridis, I., Rohland, N., Mallick, S., Patterson, N., Roodenberg, S. A., Harney, E., Stewardson, K., Fernandes, D., Novak, M., et al. (2015) · Nature · doi:10.1038/nature16152 · Cited by 1,515
Stacks 2: Analytical methods for paired‐end sequencing improve RADseq‐based population genomics
Rochette, N. C., Rivera‐Colón, A. G., & Catchen, J. (2019) · Molecular Ecology · doi:10.1111/mec.15253 · Cited by 1,265
MitoZ: a toolkit for animal mitochondrial genome assembly, annotation and visualization
Meng, G., Li, Y., Yang, C., & Liu, S. (2019) · Nucleic Acids Research · doi:10.1093/nar/gkz173 · Cited by 1,194
The genetic history of Ice Age Europe
Fu, Q., Posth, C., Hajdinjak, M., Petr, M., Mallick, S., Fernandes, D., Furtwängler, A., Haak, W., Meyer, M., Mittnik, A., et al. (2016) · Nature · doi:10.1038/nature17993 · Cited by 959
Metadata
- Authors
- Korneliussen, T. S., Albrechtsen, A., & Nielsen, R.
- Affiliations
- Natural History Museum of Denmark (DK)
- University of Copenhagen (DK)
- Corresponding author
- Korneliussen, Thorfinn Sand
- Venue
- BMC Bioinformatics
- Published
- 2014-11-25
- Volume
- 15
- Issue
- 1
- Pages
- 356-356
- ISSN
- 1471-2105 (ISSN-L)
- PMC
- PMC4248462
- OpenAlex
- W2129034036
- Cited by
- 2,982
- References
- 36
- Citations by year
- 2026: 332 · 2025: 419 · 2024: 401 · 2023: 335 · 2022: 356 · 2021: 364 · 2020: 233 · 2019: 188 · 2018: 143 · 2017: 97 · 2016: 74 · 2015: 28 · 2014: 2
- FWCI
- 27.45
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Locations
- BMC Bioinformatics (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Research at the University of Copenhagen (University of Copenhagen) (publishedVersion, cc-by)
- eScholarship (California Digital Library) (submittedVersion)
- eScholarship (California Digital Library) (submittedVersion, cc-by)
- Funders
- Villum Fonden (00007171)
- National Research Foundation
- Danmarks Grundforskningsfond (DNRF94)
- National Institutes of Health (2R14003229, 2R14003229-07)
- U.S. Public Health Service (2R14003229-07)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genetic diversity and population structure
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genetic Associations and Epidemiology
- Keywords
- next-generation sequencing data analysis, association mapping, population genetics, genotype likelihoods, multithreaded programs, summary statistics, BAM files, imputed genotypes
- MeSH terms
- Gene Frequency, Genetics, Population, Genotype, Software, Likelihood Functions, Polymorphism, Single Nucleotide, High-Throughput Nucleotide Sequencing
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
