

Molecular Pathway-Based Classification of Ectodermal Dysplasias: First Five-Yearly Update
Open accessThe DOI matches its Crossref record; 56 of the 56 references it cites were found in a registry or an index; 7 of the 16 sources its claims cite were checked for support.
Abstract
Updated molecular pathway-based classification of ectodermal dysplasias, listing 49 conditions with clarified molecular basis including 15 new entities.
Overview
CoChat Summary
## TL;DR
This paper updates the 2017 molecular pathway-based classification of ectodermal dysplasias (EDs) by reviewing recent literature and expert consensus. It reports 49 currently known EDs with clarified molecular bases, including 15 new entities, and distinguishes them from 34 syndromes with ED-like features that are likely better classified elsewhere.
## Key findings
- The updated “core ED classification” includes 49 currently known ectodermal dysplasias with clarified molecular genetic bases, of which 15 are new entities compared with the prior list.
- The authors state that the previous broader list of 97 distinct ED entities was reduced to 49 after excluding syndromes without clear genetic evidence or with insufficient/obsolete documentation.
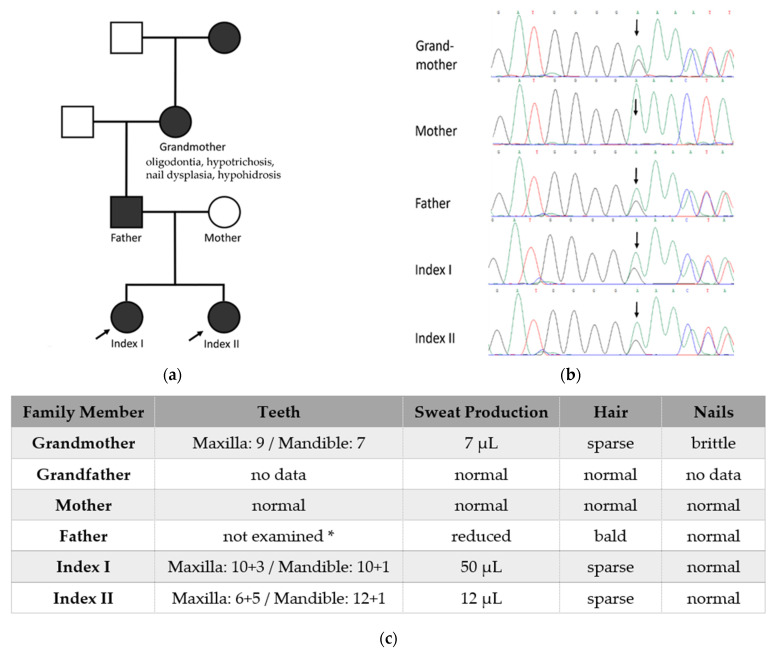
- A new ED linked to LRP6 is described in more detail; in one German family, 5 affected patients had a heterozygous frameshift mutation (c.4219dupG; p.E1407Gfs*5) that was absent in unaffected relatives.
- The paper says the authors identified 43 OMIM-listed diseases labeled as EDs without reported causative variants; they established contact for 35, received 17 responses, and found that for 6 diseases genetic material was unavailable and/or contact was impossible.
## Methods
- Literature and database update: the authors searched PubMed and OMIM for “ectodermal dysplasia,” focusing mainly on changes from the last 5 years.
- Retrospective case follow-up: for older ED entries lacking molecular confirmation, they tried to contact original authors or their institutions to determine whether patient genetic material might still be available.
- Expert review: a group of experts from the 8th International Conference on Ectodermal Dysplasias and prior panel members reviewed proposed changes and finalized the classification table.
## Limitations
- The paper explicitly notes that a five-yearly update is only a “snapshot” and will need further expansion and improvement as new data emerge.
- It states there are still “blind spots” in exome/genome testing and that the list will need regular updating.
- Evident from the text: some exclusions and reclassifications were made because older cases lacked molecular diagnosis, had only one or a few publications, or could not be genetically clarified due to loss of contact or unavailable material.
Model-written summary. Check it against the paper before relying on it.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
4 Valid · 0 Not valid · 4 Not checked · 17 Own results and interpretations
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“To be mentioned here, for example, variants of TSPEAR [47,48], HOXC13 [49] or CST6 [50] have been shown to underlie new forms of ED.”
§ Molecular Pathway-Based Classification of Ectodermal Dysplasias: First Five-Yearly Update > 5. Selection of Diseases to Be Classified > 5.2. New Inclusions
What the sources say
- Supportedcites [47] Peled, A., Sarig, O., Samuelov, L., Bertolini, M., Ziv, L., Weissglas-Volkov, D., Eskin-Schwartz, M., Adase, C.A., Malchin, N., and Bochner, R. (2016). Mutations in TSPEAR, encoding a regulator of Notch signaling, affect tooth and hair follicle morphogenesis. PLoS Genet., 12. source
The citation to this paper is attached to TSPEAR, not HOXC13 or CST6. The Abstract reports TSPEAR mutations segregating with a new form of ectodermal dysplasia in three families, and the Discussion describes that form as caused by TSPEAR mutations. (LLM, conf=0.94)
- Supportedcites [48] Bowles, 2021, TSPEAR variants are primarily associated with ectodermal dysplasia and tooth agenesis but not hearing loss: A novel cohort study, Am. J. Med. Genet. A source
The Discussion reports that ectodermal dysplasia with tooth agenesis occurred in 7 of 21 individuals with biallelic TSPEAR variants and describes substantial evidence linking TSPEAR variants to ED. This supports the TSPEAR portion of the sentence; HOXC13 and CST6 are attributed to other citations. (LLM, conf=0.91)
- Not checked yetcites [49] Lin, 2012, Loss-of-function mutations in HOXC13 cause pure hair and nail ectodermal dysplasia, Am. J. Hum. Genet. source
Checked before — its earlier verdict is held back until a new check runs.
- Partly supportedcites [50] Jansen, 2019, Deficiency of the human cysteine protease inhibitor cystatin M/E causes hypotrichosis and dry skin, Genet. Med. sourceExperimental — accuracy not yet measured
The Introduction reports that a CST6 loss-of-function variant causes a novel autosomal recessive hypotrichosis syndrome, and the Results describe affected family members with dry skin. The paper does not identify the syndrome as ectodermal dysplasia (ED); the Discussion classifies it as a heritable hair disease. This citation supports the CST6 link to a new syndrome, but not its stated ED classification. (LLM, conf=0.83)
plainstates a cause- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Therefore, the effect of this presumably dominant-negative mutation on early development may be more widespread than that of previously described pathogenic LRP6 variants [54,55].”
§ Molecular Pathway-Based Classification of Ectodermal Dysplasias: First Five-Yearly Update > 5. Selection of Diseases to Be Classified > 5.2. New Inclusions
What the sources say
- Supportedcites [54] Ockeloen, 2016, Novel mutations in LRP6 highlight the role of WNT signaling in tooth agenesis, Genet. Med. source
The citation identifies previously described pathogenic LRP6 variants. The Results report LRP6 variants considered causal for severe tooth agenesis, and the Discussion describes associated dental anomalies. The proposed comparison with the citing authors’ dominant-negative mutation is their own inference, not a finding attributed to this paper. (LLM, conf=0.87)
- Partly supportedcites [55] Lee, Y., Chae, W., Kim, Y.J., and Kim, J.-W. (2022). Novel LRP6 mutations causing non-syndromic oligodontia. J. Pers. Med., 12. sourceExperimental — accuracy not yet measured
The Discussion describes a previously reported LRP6 frameshift associated with orofacial clefting and suggests its dominant-negative effect would be more harmful than that of other frameshift mutations degraded by nonsense-mediated decay. It does not compare the citing paper’s mutation with previously described LRP6 variants or establish that its effects on early development may be more widespread. The Conclusions describe this paper’s two newly identified mutations as predicted to cause transcript degradation, not dominant-negative effects. (LLM, conf=0.84)
hedgedstates a possible cause- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
None of the sources cited for this claim have been checked yet.
In the paper
“Since the last literature search for the revised ED classification in 2017 [3], several new EDs and their genetic causes have been described.”
§ Molecular Pathway-Based Classification of Ectodermal Dysplasias: First Five-Yearly Update > 5. Selection of Diseases to Be Classified > 5.2. New Inclusions
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“We identified a total of 43 diseases in OMIM that were listed as EDs without any report about causative gene variants.”
§ Molecular Pathway-Based Classification of Ectodermal Dysplasias: First Five-Yearly Update > 5. Selection of Diseases to Be Classified > 5.1. Exclusions Many of the OMIM-listed EDs were described in the last century only based on their clinical phenotypes, and molecular diagnostics was not performed. It is well possible, however, that recently discovered variants of ED-related genes also caused diseases reported in the last century. This is why we have tried to investigate disease entries retrospectively whose genetic background had not been elucidated by the time of reporting and, if necessary, have reclassified those diseases or removed them from the list of EDs to be classified. We identified a total of 43 diseases in OMIM that were listed as EDs without any report about causative gene variants. For the majority of these EDs, no more than three publications were available, the last of which was usually older than 15 years. For example, two reports about deafness with anhidrotic ectodermal dysplasia (OMIM #125050) have been published, the most recent of which appeared in 1951 [46]. Therefore, we tried to contact the authors of these reports using the contact details indicated in the published paper. Since many of the authors are not alive anymore or retired a long time ago, direct contact could often not be established. In this case, we contacted the institution where the relevant research was conducted. We were able to establish contacts with the authors of 35 OMIM-listed diseases or their institutions and have received 17 responses to the question whether genetic material of the patients would still be available. For six diseases it became clear that genetic material is not available, nor will contact be possible with the patients and/or the authors of the publication. We are continuing this research that may allow an expansion of Table 1 in case of positive findings. The diseases investigated in this way are listed in the Supplementary Material (Table S1). Ultimately, we decided that we should not classify diseases for which the genetic cause is unknown and only one publication exists, and/or the last publication dated back more than 15 years and/or no response from the authors was received.
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“For six diseases it became clear that genetic material is not available, nor will contact be possible with the patients and/or the authors of the publication.”
§ Molecular Pathway-Based Classification of Ectodermal Dysplasias: First Five-Yearly Update > 5. Selection of Diseases to Be Classified > 5.1. Exclusions Many of the OMIM-listed EDs were described in the last century only based on their clinical phenotypes, and molecular diagnostics was not performed. It is well possible, however, that recently discovered variants of ED-related genes also caused diseases reported in the last century. This is why we have tried to investigate disease entries retrospectively whose genetic background had not been elucidated by the time of reporting and, if necessary, have reclassified those diseases or removed them from the list of EDs to be classified. We identified a total of 43 diseases in OMIM that were listed as EDs without any report about causative gene variants. For the majority of these EDs, no more than three publications were available, the last of which was usually older than 15 years. For example, two reports about deafness with anhidrotic ectodermal dysplasia (OMIM #125050) have been published, the most recent of which appeared in 1951 [46]. Therefore, we tried to contact the authors of these reports using the contact details indicated in the published paper. Since many of the authors are not alive anymore or retired a long time ago, direct contact could often not be established. In this case, we contacted the institution where the relevant research was conducted. We were able to establish contacts with the authors of 35 OMIM-listed diseases or their institutions and have received 17 responses to the question whether genetic material of the patients would still be available. For six diseases it became clear that genetic material is not available, nor will contact be possible with the patients and/or the authors of the publication. We are continuing this research that may allow an expansion of Table 1 in case of positive findings. The diseases investigated in this way are listed in the Supplementary Material (Table S1). Ultimately, we decided that we should not classify diseases for which the genetic cause is unknown and only one publication exists, and/or the last publication dated back more than 15 years and/or no response from the authors was received.
boosted- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
7 more claims
Create a free account to see all 25 claims and to verify the 2 references still open.
- It may state two separate claims as one. 8
- Probably not a substantial claim. 6
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“The most recent classification was proposed by Peschel et al.[5] as a part of the five-yearly update on EDs. This system adhered to the previous genetic scheme of classification while adding 15 new entities or genes that have been recently studied.”
From Ectodermal Dysplasia – An Overview and Update (2024) · cites this paper as [5]
Citations
Cites
CYLD is a deubiquitinating enzyme that negatively regulates NF-κB activation by TNFR family members
Trompouki, E., Hatzivassiliou, E., Tsichritzis, T., Farmer, H., Ashworth, A., & Mosialos, G. (2003) · Nature · doi:10.1038/nature01803 · Cited by 961 · cited 1×
“The termination of the signaling is possibly achieved by CYLD lysine-63 deubiquitinase which deubiquitinates the TRAF6 protein [37].”
Backgroundarrow encodes an LDL-receptor-related protein essential for Wingless signalling
Wehrli, M., Dougan, S. T., Caldwell, K., O'Keefe, L., Schwartz, S., Vaizel-Ohayon, D., Schejter, E., Tomlinson, A., & DiNardo, S. (2000) · Nature · doi:10.1038/35035110 · Cited by 871 · cited 1×
“Its gene belongs to the LDLR gene family [51].”
BackgroundKere, J., Srivastava, A. K., Montonen, O., Zonana, J., Thomas, N., Ferguson, B., Munoz, F., Morgan, D., Clarke, A., Baybayan, P., et al. (1996) · Nature Genetics · doi:10.1038/ng0895-409 · Cited by 749 · cited 5×
“The first gene discovered as being linked to ED was EDA, an X-chromosomal gene encoding the signaling protein ectodysplasin A1 (EDA1) [6].”
BackgroundInfluentialSelf-citationA Mechanism for Wnt Coreceptor Activation
Tamai, K., Zeng, X., Liu, C., Zhang, X., Harada, Y., Chang, Z., & He, X. (2004) · Molecular Cell · doi:10.1016/s1097-2765(03)00484-2 · Cited by 490 · cited 1×
“The protein has an extracellular domain, a transmembrane portion, and an intracellular part that contains five Pro-Pro-Pro-Ser/Thr-Pro (PPPS/TP) motifs [52].”
BackgroundGene defect in ectodermal dysplasia implicates a death domain adapter in development
Headon, D. J., Emmal, S. A., Ferguson, B. M., Tucker, A. S., Justice, M. J., Sharpe, P. T., Zonana, J., & Overbeek, P. A. (2001) · Nature · doi:10.1038/414913a · Cited by 378 · cited 1×
“, EDAR, coding for the EDA1 receptor, and EDARADD, encoding the adaptor protein “EDAR-associated death domain” [8].”
BackgroundEin neues Verfahren zu der klinischen Untersuchung der Schweißabsonderung
Minor, V. (1928) · Deutsche Zeitschrift für Nervenheilkunde · doi:10.1007/bf01652699 · Cited by 352 · cited 1×
“Clinical investigations of the presence of sweat glands and sweating can be performed by using a starch–iodine test [18], assessing the sweat pore density crudely with a hand lens or determining it more precisely with a confocal microscope, and/or measuring the pilocarpine-induced sweat production or the skin conducta…”
BackgroundCluzeau, C., Hadj-Rabia, S., Jambou, M., Mansour, S., Guigue, P., Masmoudi, S., Bal, E., Chassaing, N., Vincent, M.-C., Viot, G., et al. (2011) · Human Mutation · doi:10.1002/humu.21384 · Cited by 311
Self-citationEctodermal dysplasias: Classification and organization by phenotype, genotype and molecular pathway.
Wright, J. T., Fete, M., Schneider, H., Zinser, M., Koster, M. I., Clarke, A. J., Hadj‐Rabia, S., Tadini, G., Pagnan, N., Visinoni, A. F., et al. (2019) · American Journal of Medical Genetics Part A · doi:10.1002/ajmg.a.61045 · Cited by 185 · cited 7×
“On the other hand, mutated IKBKG, which is causative of Incontinentia pigmenti (OMIM #308300), leads to short stature, abnormal eye development, hypodontia, and may also affect the central nervous system [3].”
BackgroundInfluentialSelf-citationPrenatal correction of x-linked hypohidrotic ectodermal dysplasia
Schneider, H., Faschingbauer, F., Schuepbach-Mallepell, S., Körber, I., Wohlfart, S., Dick, A., Wahlbuhl, M., Kowalczyk-Quintas, C., Vigolo, M., Kirby, N., et al. (2018) · New England Journal of Medicine · doi:10.1056/nejmoa1714322 · Cited by 183
Self-citationEctodermal dysplasias: Clinical and molecular review
Visinoni, Á. F., Lisboa‐Costa, T., Pagnan, N. A. B., & Chautard‐Freire‐Maia, E. A. (2009) · American Journal of Medical Genetics Part A · doi:10.1002/ajmg.a.32864 · Cited by 181 · cited 2×
“In more severe cases, nails are hypoplastic, dystrophic or absent [30,31].”
BackgroundSelf-citationGenetic analysis: Wnt and other pathways in nonsyndromic tooth agenesis
Yu, M., Wong, S., Han, D., & Cai, T. (2019) · Oral Diseases · doi:10.1111/odi.12931 · Cited by 155 · cited 2×
“Pathogenic variants of genes involved in this pathway are known to cause either nonsyndromic tooth agenesis, hypohidrotic (or hidrotic) ED, or Goltz syndrome [39].”
BackgroundEzer, S. (1999) · Human Molecular Genetics · doi:10.1093/hmg/8.11.2079 · Cited by 143 · cited 3×
“The EDA2-EDA2R interaction induces NF-κ B activation via the non-canonical EDA/NF-κ B pathway [38], but whether pathogenic EDA2R ( XEDAR ) variants can cause HED is still under investigation [33].”
BackgroundShimomura, Y., Wajid, M., Shapiro, L., & Christiano, A. M. (2008) · Development · doi:10.1242/dev.006718 · Cited by 122 · cited 1×
“For example, cadherin-3 defects due to pathogenic CDH3 variants could be included both in the structure group and as a target of p63 in its pathway [32].”
BackgroundHo, M., Thompson, B. S., Fisk, J. N., Nebert, D. W., Bruford, E. A., Vasiliou, V., & Bunick, C. G. (2022) · Human Genomics · doi:10.1186/s40246-021-00374-9 · Cited by 109 · cited 1×
“Keratins are intermediate filament proteins and are essential for providing structural stability to hair, epidermis, and nails [43].”
BackgroundMorlon, A., Munnich, A., & Smahi, A. (2005) · Human Molecular Genetics · doi:10.1093/hmg/ddi405 · Cited by 107
Exome Sequence Identifies RIPK4 as the Bartsocas- Papas Syndrome Locus
Mitchell, K., O’Sullivan, J., Missero, C., Blair, E., Richardson, R., Anderson, B., Antonini, D., Murray, J. C., Shanske, A. L., Schutte, B. C., et al. (2011) · The American Journal of Human Genetics · doi:10.1016/j.ajhg.2011.11.013 · Cited by 101 · cited 1×
“Tumor protein p63 induces directly or indirectly receptor-interacting serine/threonine kinase 4 (RIPK4) [41] or EDAR [42].”
Backgroundp63 in Skin Development and Ectodermal Dysplasias
Koster, M. I. (2010) · Journal of Investigative Dermatology · doi:10.1038/jid.2010.119 · Cited by 100 · cited 1×
“Various human developmental disorders caused by alterations in p63 signaling clearly demonstrate the importance of p63 in the development of the skin and its appendages [40].”
BackgroundSelf-citationPrevalence and prevention of severe complications of hypohidrotic ectodermal dysplasia in infancy
Blüschke, G., Nüsken, K.-D., & Schneider, H. (2010) · Early Human Development · doi:10.1016/j.earlhumdev.2010.04.008 · Cited by 97
Self-citationPrice, J., Wright, J., Walker, S., Crawford, P. J. M., Aldred, M. J., & Hart, T. C. (1999) · Clinical Genetics · doi:10.1034/j.1399-0004.1999.550105.x · Cited by 89 · cited 1×
“Variants of the gene DLX3 should be mentioned here, which cause tricho–dento–osseous syndrome (TDO, OMIM #190320) [29].”
BackgroundSelf-citationPetrof, G., Mellerio, J. E., & McGrath, J. A. (2012) · British Journal of Dermatology · doi:10.1111/j.1365-2133.2011.10640.x · Cited by 87 · cited 1×
“Pathogenic variants of genes involved in cell–cell connection/communication are also involved in disorders of ectodermal development [44].”
BackgroundThe prevalence of X-linked hypohidrotic ectodermal dysplasia (XLHED) in Denmark, 1995–2010
Nguyen-Nielsen, M., Skovbo, S., Svaneby, D., Pedersen, L. H., & Fryzek, J. P. (2013) · European Journal of Medical Genetics · doi:10.1016/j.ejmg.2013.01.012 · Cited by 83
Loss-of-function mutations in HOXC13 cause pure hair and nail ectodermal dysplasia
Lin, Z., Chen, Q., Shi, L., Lee, M., Giehl, K. A., Tang, Z., Wang, H., Zhang, J., Yin, J., Wu, L., et al. (2012) · The American Journal of Human Genetics · doi:10.1016/j.ajhg.2012.08.029 · Cited by 79 · cited 1×
“To be mentioned here, for example, variants of TSPEAR [47,48], HOXC13 [49] or CST6 [50] have been shown to underlie new forms of ED.”
BackgroundNovel mutations in LRP6 highlight the role of WNT signaling in tooth agenesis
Ockeloen, C. W., Khandelwal, K. D., Dreesen, K., Ludwig, K. U., Sullivan, R., van Rooij, I. A. L. M., Thonissen, M., Swinnen, S., Phan, M., Conte, F., et al. (2016) · Genetics in Medicine · doi:10.1038/gim.2016.10 · Cited by 78 · cited 2×
“An explanation as to why mutated LRP6 led to ED in that family, whereas it used to be linked to non-syndromic tooth agenesis in previous publications [54,55], may arise from considering the molecular function of the protein [53].”
BackgroundOrodental manifestations in ectodermal dysplasia—A review
Bergendal, B. (2014) · American Journal of Medical Genetics Part A · doi:10.1002/ajmg.a.36571 · Cited by 71 · cited 2×
“Teeth may be relatively small, widely spaced and susceptible to mechanical damage [28].”
BackgroundEtiology and pathogenesis of ectodermal dysplasias
Itin, P. H. (2014) · American Journal of Medical Genetics Part A · doi:10.1002/ajmg.a.36550 · Cited by 69
Mutant p63 Affects Epidermal Cell Identity through Rewiring the Enhancer Landscape
Qu, J., Tanis, S. E. J., Smits, J. P. H., Kouwenhoven, E. N., Oti, M., van den Bogaard, E. H., Logie, C., Stunnenberg, H. G., van Bokhoven, H., Mulder, K. W., et al. (2018) · Cell Reports · doi:10.1016/j.celrep.2018.11.039 · Cited by 58
A costs analysis of dental treatment for ectodermal dysplasia
MURDOCK, S., LEE, J. Y., GUCKES, A., & WRIGHT, J. T. (2005) · The Journal of the American Dental Association · doi:10.14219/jada.archive.2005.0343 · Cited by 57 · cited 1×
“In some countries, a correct diagnosis can also mitigate the enormous socio-economic burden associated with an ED through, for example, adequate dental insurance [10].”
BackgroundSelf-citationWNT10A , dermatology and dentistry
Doolan, B. J., Onoufriadis, A., Kantaputra, P., & McGrath, J. A. (2021) · British Journal of Dermatology · doi:10.1111/bjd.20601 · Cited by 56 · cited 2×
“In humans, 19 different Wnt ligands have been described, of which Wnt-10A and Wnt-10B are clinically the most important ones for tooth development [34].”
BackgroundPeled, A., Sarig, O., Samuelov, L., Bertolini, M., Ziv, L., Weissglas-Volkov, D., Eskin-Schwartz, M., Adase, C. A., Malchin, N., Bochner, R., et al. (2016) · PLOS Genetics · doi:10.1371/journal.pgen.1006369 · Cited by 55 · cited 2×
“In addition to the EDA/NF-κ B pathway, the p63 transcription factor pathway, and the Wnt/ β -catenin pathway, we included a group of genes that play a role in the development of cell structure or keratinization, and, as a fifth group, EDs related to other pathways, such as Notch signaling (e.g., TSPEAR variants).”
BackgroundSweating ability and genotype in individuals with X-linked hypohidrotic ectodermal dysplasia
Schneider, H., Hammersen, J., Preisler-Adams, S., Huttner, K., Rascher, W., & Bohring, A. (2011) · Journal of Medical Genetics · doi:10.1136/jmg.2010.084012 · Cited by 54 · cited 1×
“Clinical investigations of the presence of sweat glands and sweating can be performed by using a starch–iodine test [18], assessing the sweat pore density crudely with a hand lens or determining it more precisely with a confocal microscope, and/or measuring the pilocarpine-induced sweat production or the skin conducta…”
BackgroundSelf-citationEctodermal Dysplasias: A Clinical and Molecular Review
García‐Martín, P., Hernández‐Martín, A., & Torrelo, A. (2013) · Actas Dermo-Sifiliográficas · doi:10.1016/j.adengl.2013.05.001 · Cited by 53 · cited 1×
“In more severe cases, nails are hypoplastic, dystrophic or absent [30,31].”
BackgroundUpdate on ectodermal dysplasias clinical classification
Pagnan, N. A. B., & Visinoni, Á. F. (2014) · American Journal of Medical Genetics Part A · doi:10.1002/ajmg.a.36616 · Cited by 52 · cited 3×
“Classification of Ectodermal Dysplasias and Their Correct Diagnosis In the latest approach to a clinical, phenotype-based classification of EDs, the relevant disease entities were divided up into 11 subgroups according to the structures affected [2].”
BackgroundInfluentialSelf-citationXEDAR activates the non-canonical NF-κB pathway
Verhelst, K., Gardam, S., Borghi, A., Kreike, M., Carpentier, I., & Beyaert, R. (2015) · Biochemical and Biophysical Research Communications · doi:10.1016/j.bbrc.2015.08.019 · Cited by 42 · cited 1×
“The EDA2-EDA2R interaction induces NF-κB activation via the non-canonical EDA/NF-κB pathway [38], but whether pathogenic EDA2R (XEDAR) variants can cause HED is still under investigation [33].”
BackgroundDisplasias ectodérmicas: revisión clínica y molecular
García-Martín, P., Hernández-Martín, A., & Torrelo, A. (2013) · Actas Dermo-Sifiliográficas · doi:10.1016/j.ad.2012.07.012 · Cited by 41
Wohlfart, S., Hammersen, J., & Schneider, H. (2016) · Journal of Human Genetics · doi:10.1038/jhg.2016.75 · Cited by 41 · cited 2×
“The EDA2-EDA2R interaction induces NF-κB activation via the non-canonical EDA/NF-κB pathway [38], but whether pathogenic EDA2R (XEDAR) variants can cause HED is still under investigation [33].”
BackgroundSelf-citationLrp6 Dynamic Expression in Tooth Development and Mutations in Oligodontia
Yu, M., Fan, Z., Wong, S. W., Sun, K., Zhang, L., Liu, H., Feng, H., Liu, Y., & Han, D. (2021) · Journal of Dental Research · doi:10.1177/0022034520970459 · Cited by 41 · cited 14×
“Pathogenic variants of LRP6 have been known so far only as being causative for non-syndromic tooth agenesis (OMIM #616724).”
BackgroundResultInfluentialSalivary gland function in persons with ectodermal dysplasias
Nordgarden, H., Storhaug, K., Lyngstadaas, S. P., & Jensen, J. L. (2003) · European Journal of Oral Sciences · doi:10.1034/j.1600-0722.2003.00058.x · Cited by 38 · cited 1×
“Decreased secretion of salivary glands can cause a dry mouth and nutritional problems, alter the quality of the voice [22], or increase the susceptibility to opportunistic oral infections [23].”
BackgroundFunctional studies for the TRAF6 mutation associated with hypohidrotic ectodermal dysplasia
Fujikawa, H., Farooq, M., Fujimoto, A., Ito, M., & Shimomura, Y. (2013) · British Journal of Dermatology · doi:10.1111/bjd.12018 · Cited by 26 · cited 5×
“EDARADD further builds a complex with TNF-receptor-associated factor 6 (TRAF6), TAK1-binding protein 2 (TAB2) and TGF-β -activated kinase 1 (TAK1) [36].”
BackgroundInfluentialA finger on the pulse of Wnt receptor signaling
MacDonald, B. T., & He, X. (2012) · Cell Research · doi:10.1038/cr.2012.91 · Cited by 25 · cited 2×
“An explanation as to why mutated LRP6 led to ED in that family, whereas it used to be linked to non-syndromic tooth agenesis in previous publications [54,55], may arise from considering the molecular function of the protein [53].”
BackgroundOphthalmic Manifestations in Patients with Ectodermal Dysplasia Syndromes
Keklíkçí, U., Yavuz, İ., Tunik, S., Baskan, Z., & Akdeniz, S. (2014) · Advances in Clinical and Experimental Medicine · doi:10.17219/acem/37235 · Cited by 25 · cited 1×
“Disorders of the tear film related to missing meibomian glands and underdeveloped lacrimal glands often lead to dry eyes and may be investigated, for example, with a Schirmer test [21].”
BackgroundLEF1 haploinsufficiency causes ectodermal dysplasia
Lévy, J., Capri, Y., Rachid, M., Dupont, C., Vermeesch, J. R., Devriendt, K., Verloes, A., Tabet, A., & Bailleul‐Forestier, I. (2020) · Clinical Genetics · doi:10.1111/cge.13714 · Cited by 25
Hypohidrotic Ectodermal Dysplasia: Breastfeeding Complications Due to Impaired Breast Development
Wahlbuhl-Becker, M., Faschingbauer, F., Beckmann, M. W., & Schneider, H. (2017) · Geburtshilfe und Frauenheilkunde · doi:10.1055/s-0043-100106 · Cited by 24
Self-citationInfluence of Reduced Saliva Production on Phonation in Patients With Ectodermal Dysplasia
Semmler, M., Kniesburges, S., Pelka, F., Ensthaler, M., Wendler, O., & Schützenberger, A. (2021) · Journal of Voice · doi:10.1016/j.jvoice.2021.06.016 · Cited by 22 · cited 1×
“Decreased secretion of salivary glands can cause a dry mouth and nutritional problems, alter the quality of the voice [22], or increase the susceptibility to opportunistic oral infections [23].”
BackgroundUpdate of recent findings in genetic hair disorders
Hayashi, R., & Shimomura, Y. (2022) · The Journal of Dermatology · doi:10.1111/1346-8138.16204 · Cited by 22 · cited 1×
“Eyebrows and eyelashes are also sparse or absent [25].”
BackgroundBowles, B., Ferrer, A., Nishimura, C. J., Pinto e Vairo, F., Rey, T., Leheup, B., Sullivan, J., Schoch, K., Stong, N., Agolini, E., et al. (2021) · American Journal of Medical Genetics Part A · doi:10.1002/ajmg.a.62347 · Cited by 20 · cited 2×
“In addition to the EDA/NF-κ B pathway, the p63 transcription factor pathway, and the Wnt/ β -catenin pathway, we included a group of genes that play a role in the development of cell structure or keratinization, and, as a fifth group, EDs related to other pathways, such as Notch signaling (e.g., TSPEAR variants).”
BackgroundNovel mutations inGJB6andGJB2in Clouston syndrome
Liu, Y. T., Guo, K., Li, J., Liu, Y., Zeng, W. H., & Geng, S. (2015) · Clinical and Experimental Dermatology · doi:10.1111/ced.12654 · Cited by 19 · cited 1×
“alterations in connexins, nectins, cadherins, desmoplakin or plakoglobin can be responsible for EDs [45].”
BackgroundCharacterization of EDARADD gene mutations responsible for hypohidrotic ectodermal dysplasia
Asano, N., Yasuno, S., Hayashi, R., & Shimomura, Y. (2021) · The Journal of Dermatology · doi:10.1111/1346-8138.16044 · Cited by 19 · cited 5×
“EDARADD further builds a complex with TNF-receptor-associated factor 6 (TRAF6), TAK1-binding protein 2 (TAB2) and TGF-β -activated kinase 1 (TAK1) [36].”
BackgroundInfluentialKaercher, T., Dietz, J., Jacobi, C., Berz, R., & Schneider, H. (2014) · Current Eye Research · doi:10.3109/02713683.2014.967869 · Cited by 18 · cited 1×
“Apart from sweat glands, other eccrine glands of ectodermal origin can also malfunction or be absent like the lacrimal, salivary, mammary, sebaceous, and meibomian glands [20].”
BackgroundSelf-citationDeficiency of the human cysteine protease inhibitor cystatin M/E causes hypotrichosis and dry skin
van den Bogaard, E. H. J., van Geel, M., van Vlijmen-Willems, I. M. J. J., Jansen, P. A. M., Peppelman, M., van Erp, P. E. J., Atalay, S., Venselaar, H., Simon, M. E. H., Joosten, M., et al. (2019) · Genetics in Medicine · doi:10.1038/s41436-018-0355-3 · Cited by 18
Thermoregulation in Ectodermal Dysplasia: A Case Series
Massey, H. C., House, J. R., & Tipton, M. (2019) · International Journal of Environmental Research and Public Health · doi:10.3390/ijerph16224514 · Cited by 18 · cited 1×
“This also applies to fever related to infections or childhood illnesses [16].”
BackgroundMajor hereditary ectodermal dysplasia
Ellingson, R. J. (1951) · The Journal of Pediatrics · doi:10.1016/s0022-3476(51)80108-2 · Cited by 14 · cited 1×
“For example, two reports about deafness with anhidrotic ectodermal dysplasia (OMIM #125050) have been published, the most recent of which appeared in 1951 [46].”
BackgroundDufour, W., Alawbathani, S., Jourdain, A.-S., Asif, M., Baujat, G., Becker, C., Budde, B., Gallacher, L., Georgomanolis, T., Ghoumid, J., et al. (2022) · Genetics in Medicine · doi:10.1016/j.gim.2022.04.022 · Cited by 14 · cited 4×
“, TRAF6 [11,12], LEF1 [13,14], or LRP6 [15].”
BackgroundInfluentialScanning Electron Microscopic Hair Shaft Analysis in Ectodermal Dysplasia Syndromes
Hirano, S. A., Reed, A. M., Rowan, B. J., Sorrells, T. C., Williams, J. V., Pariser, D. M., Hood, A. F., & Salkey, K. S. (2015) · Pediatric Dermatology · doi:10.1111/pde.12674 · Cited by 9
Peña‐Romero, A. G., Sáez‐de‐Ocariz, M., Toussaint‐Caire, S., Morán‐Villaseñor, E., Orozco‐Covarrubias, L., & Durán‐McKinster, C. (2020) · Pediatric Dermatology · doi:10.1111/pde.14415 · Cited by 9
Freire-Maia, N. (1977) · Human Heredity · doi:10.1159/000152876 · Cited by 4
Novel LRP6 Mutations Causing Non-Syndromic Oligodontia
Lee, Y., Chae, W., Kim, Y. J., & Kim, J.-W. (2022) · Journal of Personalized Medicine · doi:10.3390/jpm12091401 · Cited by 4 · cited 2×
“An explanation as to why mutated LRP6 led to ED in that family, whereas it used to be linked to non-syndromic tooth agenesis in previous publications [54,55], may arise from considering the molecular function of the protein [53].”
Background
Cited by
Enamel Phenotypes: Genetic and Environmental Determinants
Wright, J. T. (2023) · Genes · doi:10.3390/genes14030545 · Cited by 81
“There are many examples of genes associated with enamel defects that are also causative of pathology in other tissues such as skin and hair [41].”
BackgroundSelf-citationSchneider, H., Schweikl, C., Faschingbauer, F., Hadj-Rabia, S., & Schneider, P. (2023) · International Journal of Molecular Sciences · doi:10.3390/ijms24087155 · Cited by 45
“Ectodermal dysplasias are congenital genetic conditions affecting the development and/or homeostasis of two or more derivatives from the embryonic ectoderm, such as hair, teeth, nails, and certain glands in the skin and mucous membranes [1,2].”
BackgroundSelf-citationEctodermal dysplasias: New perspectives on the treatment of so far immedicable genetic disorders
Schneider, H. (2022) · Frontiers in Genetics · doi:10.3389/fgene.2022.1000744 · Cited by 26
Self-citationEctodermal Dysplasia – An Overview and Update
Dev, A., Malhi, K., & Mahajan, R. (2024) · Indian Dermatology Online Journal · doi:10.4103/idoj.idoj_599_23 · Cited by 19
“[5] as a part of the five‑yearly update on EDs.”
BackgroundPrevalence and Patient Characteristics of Ectodermal Dysplasias in Denmark
Herlin, L. K., Schmidt, S. A. J., Hermann, X. B., Rønholt, K., Bygum, A., Schuster, A., Lei, U., Mogensen, M., Vinding, G. R., Djursby, M., et al. (2024) · JAMA Dermatology · doi:10.1001/jamadermatol.2024.0036 · Cited by 14
Metadata
- Authors
- Peschel, N., Wright, J. T., Koster, M. I., Clarke, A. J., Tadini, G., Fete, M., Hadj-Rabia, S., Sybert, V. P., Norderyd, J., Maier-Wohlfart, S., Fete, T. J., Pagnan, N., Visinoni, A. F., & Schneider, H.
- Affiliations
- Friedrich-Alexander-Universität Erlangen-Nürnberg (DE)
- University of North Carolina at Chapel Hill (US)
- East Carolina University (US)
- Cardiff University (GB)
- University of Milan (IT)
- National Foundation for Ectodermal Dysplasias (US)
- Hôpital Necker-Enfants Malades (FR)
- University of Washington (US)
- Jönköping University (SE)
- Center for Ectodermal Dysplasias, University Hospital Erlangen, 91054 Erlangen, Germany
- University of Missouri (US)
- Universidade Federal do Paraná (BR)
- School of Health Sciences, Positivo University, Curitiba 81290-000, PR, Brazil
- Corresponding author
- Peschel, Nicolai
- ORCID
- Venue
- Genes
- Publisher
- MDPI AG
- Published
- 2022-12-10
- Volume
- 13
- Issue
- 12
- Pages
- 2327
- ISSN
- 2073-4425 (ISSN-L)
- PMC
- PMC9778228
- OpenAlex
- W4312125535
- Cited by
- 60
- References
- 56
- Citations by year
- 2026: 12 · 2025: 18 · 2024: 23 · 2023: 5 · 2022: 2
- FWCI
- 3.92
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- Locations
- Genes (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- ORCA Online Research @Cardiff (Cardiff University) (acceptedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- OPUS Repository (Kooperativer Bibliotheksverbund Berlin-Brandenburg) (submittedVersion, cc-by)
- Funders
- Deutsche Forschungsgemeinschaft and Friedrich-Alexander-Universität Erlangen-Nürnberg
- Topics
- Health Sciences › Dentistry › General Dentistry › dental development and anomalies
- Health Sciences › Medicine › Dermatology › Genetic and rare skin diseases.
- Health Sciences › Medicine › Dermatology › Hair Growth and Disorders
- Keywords
- ectodermal dysplasia, rare diseases, LRP6, ectodermal development, rare disease diagnosis
- MeSH terms
- Ectodermal Dysplasia, Humans, Syndrome, Rare Diseases, PubMed
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
The claim verdicts this report counts are experimental: their accuracy is not yet measured.
A language model drew the claims from the paper on 2026-10-10, under version 3 of the claim definition (model deepseek-v4.1-flash, prompt extract-v3).
A second model labelled each claim on 2026-10-10 (model jev-1.13-20260917, definition revision gold-guideline-v0.2).
Cited sources are checked with a language model’s help (gpt-6-sol).
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
Data sources
Bibliographic data: Semantic Scholar, OpenAlex, Crossref.
Contains information from the Semantic Scholar Academic Graph Datasets, made available under the ODC Attribution License (ODC-BY 1.0).
Semantic Scholar’s data is described in: Kinney, R., et al. (2023). The Semantic Scholar Open Data Platform. arXiv:2301.10140.
