

The DOI matches its Crossref record; 55 of the 55 references it cites were found in a registry or an index.
Quantum-chemical insights from deep tensor neural networks
Open accessAbstract
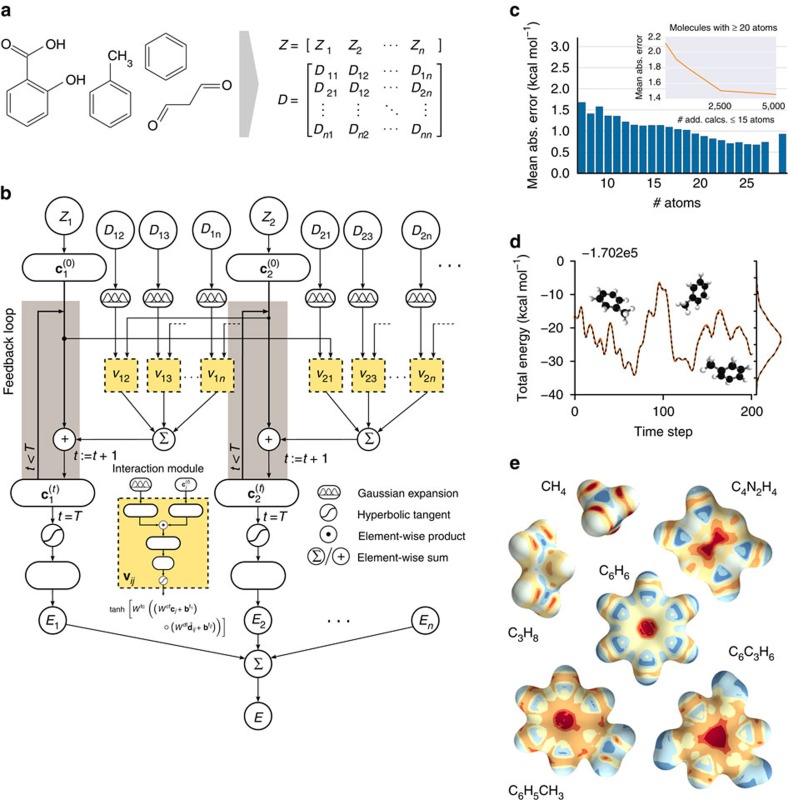
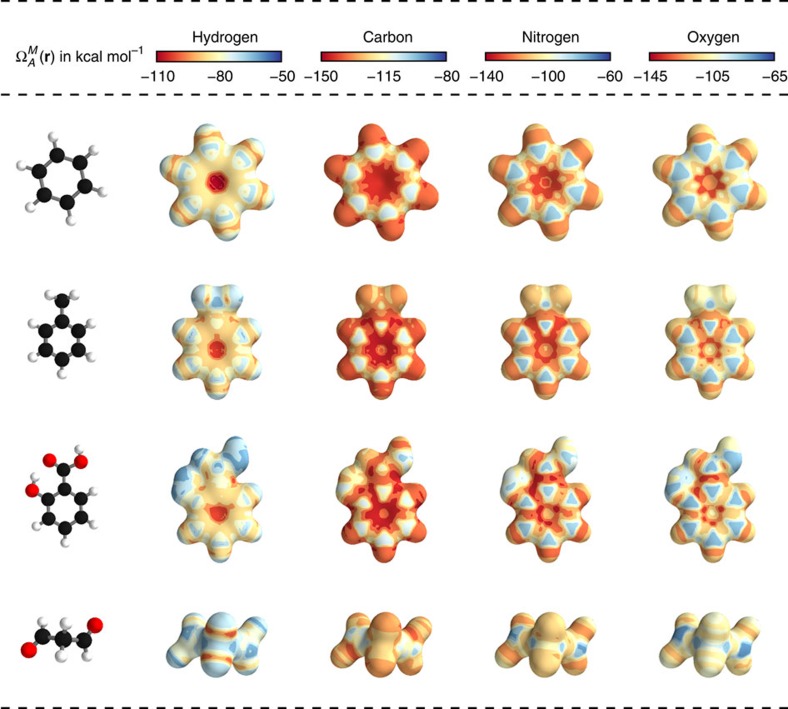
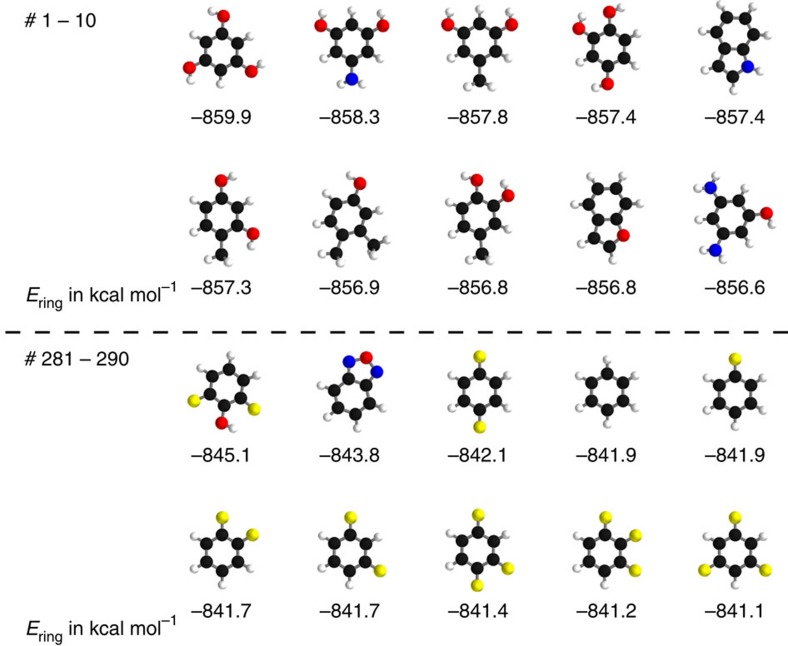
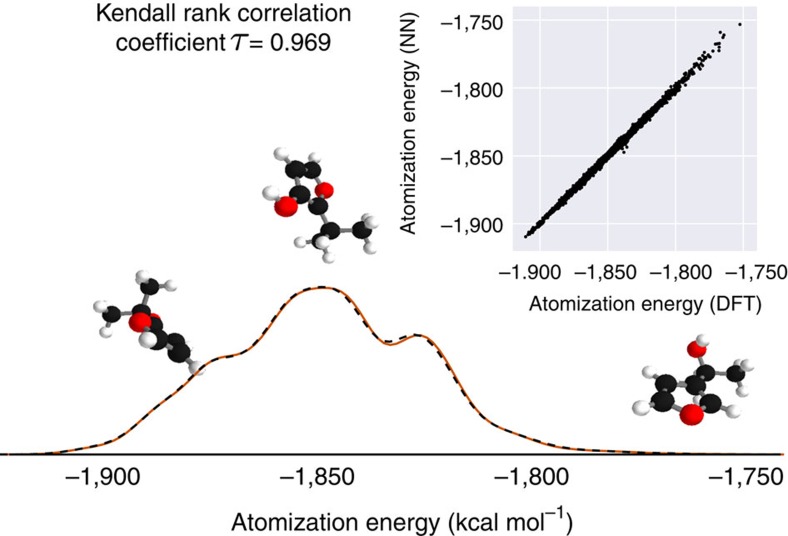
Learning from data has led to paradigm shifts in a multitude of disciplines, including web, text and image search, speech recognition, as well as bioinformatics. Can machine learning enable similar breakthroughs in understanding quantum many-body systems? Here we develop an efficient deep learning approach that enables spatially and chemically resolved insights into quantum-mechanical observables of molecular systems. We unify concepts from many-body Hamiltonians with purpose-designed deep tensor neural networks, which leads to size-extensive and uniformly accurate (1 kcal mol −1 ) predictions in compositional and configurational chemical space for molecules of intermediate size. As an example of chemical relevance, the model reveals a classification of aromatic rings with respect to their stability. Further applications of our model for predicting atomic energies and local chemical potentials in molecules, reliable isomer energies, and molecules with peculiar electronic structure demonstrate the potential of machine learning for revealing insights into complex quantum-chemical systems.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Generalized Gradient Approximation Made Simple
Perdew, J. P., Burke, K., & Ernzerhof, M. (1996) · Physical Review Letters · doi:10.1103/physrevlett.77.3865 · Cited by 215,824
ImageNet classification with deep convolutional neural networks
Krizhevsky, A., Sutskever, I., & Hinton, G. E. (2017) · Communications of the ACM · doi:10.1145/3065386 · Cited by 112,155
Density-functional thermochemistry. III. The role of exact exchange
Becke, A. D. (1993) · The Journal of Chemical Physics · doi:10.1063/1.464913 · Cited by 103,767
Lee, C., Yang, W., & Parr, R. G. (1988) · Physical Review B · doi:10.1103/physrevb.37.785 · Cited by 100,874
LeCun, Y., Bengio, Y., & Hinton, G. (2015) · Nature · doi:10.1038/nature14539 · Cited by 84,126
Density-functional exchange-energy approximation with correct asymptotic behavior
Becke, A. D. (1988) · Physical Review A · doi:10.1103/physreva.38.3098 · Cited by 54,100
Hohenberg, P., & Kohn, W. (1964) · Physical Review · doi:10.1103/physrev.136.b864 · Cited by 53,541
Stephens, P. J., Devlin, F. J., Chabalowski, C. F., & Frisch, M. J. (1994) · The Journal of Physical Chemistry · doi:10.1021/j100096a001 · Cited by 23,329
Vosko, S. H., Wilk, L., & Nusair, M. (1980) · Canadian Journal of Physics · doi:10.1139/p80-159 · Cited by 20,807
(2010) · Cited by 14,134
Cited by
Sorted by citations
A Comprehensive Survey on Graph Neural Networks
Wu, Z., Pan, S., Chen, F., Long, G., Zhang, C., & Yu, P. S. (2021) · IEEE Transactions on Neural Networks and Learning Systems · doi:10.1109/tnnls.2020.2978386 · Cited by 11,000
Graph neural networks: A review of methods and applications
Zhou, J., Cui, G., Hu, S., Zhang, Z., Yang, C., Liu, Z., Wang, L., Li, C., & Sun, M. (2020) · AI Open · doi:10.1016/j.aiopen.2021.01.001 · Cited by 6,880
Deep learning and process understanding for data-driven Earth system science
Reichstein, M., Camps-Valls, G., Stevens, B., Jung, M., Denzler, J., Carvalhais, N., & Prabhat (2019) · Nature · doi:10.1038/s41586-019-0912-1 · Cited by 5,673
Neural Message Passing for Quantum ChemistryPreprint
Gilmer, J., Schoenholz, S. S., Riley, P. F., Vinyals, O., & Dahl, G. E. (2017) · arXiv (Cornell University) · doi:10.48550/arxiv.1704.01212 · Cited by 5,678
MoleculeNet: a benchmark for molecular machine learning
Wu, Z., Ramsundar, B., Feinberg, E. N., Gomes, J., Geniesse, C., Pappu, A. S., Leswing, K., & Pande, V. (2018) · Chemical Science · doi:10.1039/c7sc02664a · Cited by 3,107
Automatic Chemical Design Using a Data-Driven Continuous Representation of Molecules
Gómez-Bombarelli, R., Wei, J. N., Duvenaud, D., Hernández-Lobato, J. M., Sánchez-Lengeling, B., Sheberla, D., Aguilera-Iparraguirre, J., Hirzel, T. D., Adams, R. P., & Aspuru-Guzik, A. (2018) · ACS Central Science · doi:10.1021/acscentsci.7b00572 · Cited by 3,106
Methods for interpreting and understanding deep neural networks
Montavon, G., Samek, W., & Müller, K.-R. (2018) · Digital Signal Processing · doi:10.1016/j.dsp.2017.10.011 · Cited by 2,665
Self-citationRecent advances and applications of machine learning in solid-state materials science
Schmidt, J., Marques, M. R. G., Botti, S., & Marques, M. A. L. (2019) · npj Computational Materials · doi:10.1038/s41524-019-0221-0 · Cited by 2,786
A Survey on Explainable Artificial Intelligence (XAI): Toward Medical XAI
Tjoa, E., & Guan, C. (2021) · IEEE Transactions on Neural Networks and Learning Systems · doi:10.1109/tnnls.2020.3027314 · Cited by 2,524
Deep Potential Molecular Dynamics: A Scalable Model with the Accuracy of Quantum Mechanics
Zhang, L., Han, J., Wang, H., Car, R., & Weinan, E. (2018) · Physical Review Letters · doi:10.1103/physrevlett.120.143001 · Cited by 2,392
SchNet – A deep learning architecture for molecules and materials
Schütt, K. T., Sauceda, H. E., Kindermans, P.-J., Tkatchenko, A., & Müller, K.-R. (2018) · The Journal of Chemical Physics · doi:10.1063/1.5019779 · Cited by 2,217
Self-citationWang, H., Zhang, L., Han, J., & Weinan, E. (2018) · Computer Physics Communications · doi:10.1016/j.cpc.2018.03.016 · Cited by 2,213
Inverse molecular design using machine learning: Generative models for matter engineering
Sánchez-Lengeling, B., & Aspuru‐Guzik, A. (2018) · Science · doi:10.1126/science.aat2663 · Cited by 2,177
E(3)-equivariant graph neural networks for data-efficient and accurate interatomic potentials
Batzner, S., Musaelian, A., Sun, L., Geiger, M., Mailoa, J. P., Kornbluth, M., Molinari, N., Smidt, T. E., & Kozinsky, B. (2022) · Nature Communications · doi:10.1038/s41467-022-29939-5 · Cited by 2,285
Analyzing Learned Molecular Representations for Property Prediction
Yang, K., Swanson, K., Jin, W., Coley, C., Eiden, P., Gao, H., Guzman-Perez, A., Hopper, T., Kelley, B., Mathea, M., et al. (2019) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.9b00237 · Cited by 1,923
Deep learning in environmental remote sensing: Achievements and challenges
Yuan, Q., Shen, H., Li, T., Li, Z., Li, S., Jiang, Y., Xu, H., Tan, W., Yang, Q., Wang, J., et al. (2020) · Remote Sensing of Environment · doi:10.1016/j.rse.2020.111716 · Cited by 1,697
Unke, O. T., Chmiela, S., Sauceda, H. E., Gastegger, M., Poltavsky, I., Schütt, K. T., Tkatchenko, A., & Müller, K.-R. (2021) · Chemical Reviews · doi:10.1021/acs.chemrev.0c01111 · Cited by 1,647
Self-citationGraph Networks as a Universal Machine Learning Framework for Molecules and Crystals
Chen, C., Ye, W., Zuo, Y., Zheng, C., & Ong, S. P. (2019) · Chemistry of Materials · doi:10.1021/acs.chemmater.9b01294 · Cited by 1,552
Explaining Deep Neural Networks and Beyond: A Review of Methods and Applications
Samek, W., Montavon, G., Lapuschkin, S., Anders, C. J., & Muller, K.-R. (2021) · Proceedings of the IEEE · doi:10.1109/jproc.2021.3060483 · Cited by 1,477
“At the same time, these methods were able to handle different types of data, including images [13], speech [20], text [10], [38], and structured data, such as molecules [162], [165] or genetic sequences [191].”
BackgroundMethodInfluentialSelf-citationShen, C. (2018) · Water Resources Research · doi:10.1029/2018wr022643 · Cited by 1,522
Deep reinforcement learning for de novo drug design
Popova, M., Isayev, O., & Tropsha, A. (2018) · Science Advances · doi:10.1126/sciadv.aap7885 · Cited by 1,433
Matminer: An open source toolkit for materials data mining
Ward, L. T., Dunn, A., Faghaninia, A., Zimmermann, N., Bajaj, S., Wang, Q. S., Montoya, J. H., Chen, J., Bystrom, K., Dylla, M. T., et al. (2018) · Computational Materials Science · doi:10.1016/j.commatsci.2018.05.018 · Cited by 1,145
Xiong, Z., Wang, D., Liu, X., Zhong, F., Wan, X., Li, X., Li, Z. S., Luo, X., Chen, K., Jiang, H., et al. (2019) · Journal of Medicinal Chemistry · doi:10.1021/acs.jmedchem.9b00959 · Cited by 1,126
Samek, W., Wiegand, T., & Müller, K.-R. (2017) · arXiv · doi:10.48550/arxiv.1708.08296 · Cited by 1,080
Self-citationA Unifying Review of Deep and Shallow Anomaly Detection
Ruff, L., Kauffmann, J., Vandermeulen, R. A., Montavon, G., Samek, W., Kloft, M., Dietterich, T. G., & Muller, K.-R. (2021) · Proceedings of the IEEE · doi:10.1109/jproc.2021.3052449 · Cited by 1,184
PhysNet: A Neural Network for Predicting Energies, Forces, Dipole Moments, and Partial Charges
Unke, O. T., & Meuwly, M. (2019) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.9b00181 · Cited by 1,131
Machine learning in materials science
Wei, J., Chu, X., Sun, X., Xu, K., Deng, H., Chen, J., Wei, Z., & Lei, M. (2019) · InfoMat · doi:10.1002/inf2.12028 · Cited by 1,069
Machine Learning Interatomic Potentials as Emerging Tools for Materials Science
Deringer, V. L., Caro, M. A., & Cśanyi, G. (2019) · Advanced Materials · doi:10.1002/adma.201902765 · Cited by 1,136
Unmasking Clever Hans predictors and assessing what machines really learn
Lapuschkin, S., Wäldchen, S., Binder, A., Montavon, G., Samek, W., & Müller, K.-R. (2019) · Nature Communications · doi:10.1038/s41467-019-08987-4 · Cited by 1,047
Self-citationExplainable machine learning for scientific insights and discoveries
Roscher, R., Bohn, B., Duarte, M. F., & Garcke, J. (2020) · IEEE Access · doi:10.1109/access.2020.2976199 · Cited by 1,040
A universal graph deep learning interatomic potential for the periodic table
Chen, C., & Ong, S. P. (2022) · Nature Computational Science · doi:10.1038/s43588-022-00349-3 · Cited by 1,140
Model Compression and Hardware Acceleration for Neural Networks: A Comprehensive Survey
Deng, B. L., Li, G., Han, S., Shi, L., & Xie, Y. (2020) · Proceedings of the IEEE · doi:10.1109/jproc.2020.2976475 · Cited by 996
Machine Learning for Molecular Simulation
Noé, F., Tkatchenko, A., Müller, K.-R., & Clementi, C. (2020) · Annual Review of Physical Chemistry · doi:10.1146/annurev-physchem-042018-052331 · Cited by 956
Self-citationDeep-Learning-Enabled On-Demand Design of Chiral Metamaterials
Ma, W., Cheng, F., & Liu, Y. (2018) · ACS Nano · doi:10.1021/acsnano.8b03569 · Cited by 928
Self-referencing embedded strings (SELFIES): A 100% robust molecular string representation
Krenn, M., Häse, F., Nigam, A., Friederich, P., & Aspuru‐Guzik, A. (2020) · Machine Learning Science and Technology · doi:10.1088/2632-2153/aba947 · Cited by 1,048
Muratov, E. N., Bajorath, J., Sheridan, R. P., Tetko, I. V., Filimonov, D., Poroikov, V., Oprea, T. I., Baskin, I. I., Varnek, A., Roitberg, A., et al. (2020) · Chemical Society Reviews · doi:10.1039/d0cs00098a · Cited by 913
Four Generations of High-Dimensional Neural Network Potentials
Behler, J. (2021) · Chemical Reviews · doi:10.1021/acs.chemrev.0c00868 · Cited by 942
DScribe: Library of descriptors for machine learning in materials science
Himanen, L., Jäger, M. O. J., Morooka, E. V., Canova, F. F., Ranawat, Y. S., Gao, D. Z., Rinke, P., & Foster, A. S. (2019) · Computer Physics Communications · doi:10.1016/j.cpc.2019.106949 · Cited by 901
Combining Machine Learning and Computational Chemistry for Predictive Insights Into Chemical Systems
Keith, J. A., Vassilev-Galindo, V., Cheng, B., Chmiela, S., Gastegger, M., Müller, K.-R., & Tkatchenko, A. (2021) · Chemical Reviews · doi:10.1021/acs.chemrev.1c00107 · Cited by 849
Self-citationLayer-Wise Relevance Propagation: An Overview
Montavon, G., Binder, A., Lapuschkin, S., Samek, W., & Müller, K. (2019) · Lecture notes in computer science · doi:10.1007/978-3-030-28954-6_10 · Cited by 883
Self-citationLess is more: Sampling chemical space with active learning
Smith, J. S., Nebgen, B., Lubbers, N., Isayev, O., & Roitberg, A. E. (2018) · The Journal of Chemical Physics · doi:10.1063/1.5023802 · Cited by 811
Data‐Driven Materials Science: Status, Challenges, and Perspectives
Himanen, L., Geurts, A., Foster, A. S., & Rinke, P. (2019) · Advanced Science · doi:10.1002/advs.201900808 · Cited by 808
Atomistic Line Graph Neural Network for improved materials property predictions
Choudhary, K. K., & DeCost, B. (2021) · npj Computational Materials · doi:10.1038/s41524-021-00650-1 · Cited by 843
Machine learning unifies the modeling of materials and molecules
Bartók, A. P., De, S., Poelking, C., Bernstein, N., Kermode, J. R., Csányi, G., & Ceriotti, M. (2017) · Science Advances · doi:10.1126/sciadv.1701816 · Cited by 764
Towards exact molecular dynamics simulations with machine-learned force fields
Chmiela, S., Sauceda, H. E., Müller, K.-R., & Tkatchenko, A. (2018) · Nature Communications · doi:10.1038/s41467-018-06169-2 · Cited by 804
Self-citationTensor field networks: Rotation- and translation-equivariant neural networks for 3D point clouds
Thomas, N., Smidt, T., Kearnes, S., Yang, L., Li, L., Kohlhoff, K., & Riley, P. (2018) · arXiv · doi:10.48550/arxiv.1802.08219 · Cited by 1,303
Learning local equivariant representations for large-scale atomistic dynamics
Musaelian, A., Batzner, S., Johansson, A., Sun, L., Owen, C. J., Kornbluth, M., & Kozinsky, B. (2023) · Nature Communications · doi:10.1038/s41467-023-36329-y · Cited by 847
Bypassing the Kohn-Sham equations with machine learning
Brockherde, F., Vogt-Maranto, L., Li, L., Tuckerman, M. E., Burke, K., & Müller, K.-R. (2017) · Nature Communications · doi:10.1038/s41467-017-00839-3 · Cited by 735
Self-citationHart, G. L. W., Mueller, T., Toher, C., & Curtarolo, S. (2021) · Nature Reviews Materials · doi:10.1038/s41578-021-00340-w · Cited by 703
Tsubaki, M., Tomii, K., & Sese, J. (2019) · Bioinformatics · doi:10.1093/bioinformatics/bty535 · Cited by 688
Smith, J. S., Nebgen, B. T., Zubatyuk, R. I., Lubbers, N., Devereux, C., Barros, K. M., Tretiak, S., Isayev, O., & Roitberg, A. E. (2019) · Nature Communications · doi:10.1038/s41467-019-10827-4 · Cited by 702
Prediction Errors of Molecular Machine Learning Models Lower than Hybrid DFT Error
Faber, F. A., Hutchison, L., Huang, B., Gilmer, J., Schoenholz, S. S., Dahl, G. E., Vinyals, O., Kearnes, S., Riley, P. F., & von Lilienfeld, O. A. (2017) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.7b00577 · Cited by 651
Towards Explainable Artificial Intelligence
Samek, W., & Müller, K. (2019) · Lecture notes in computer science · doi:10.1007/978-3-030-28954-6_1 · Cited by 704
Self-citationGraph Convolutional Policy Network for Goal-Directed Molecular Graph GenerationPreprint
You, J., Liu, B., Ying, R., Pande, V. S., & Leskovec, J. (2018) · arXiv (Cornell University) · doi:10.48550/arxiv.1806.02473 · Cited by 688
DeePMD-kit v2: A software package for deep potential models
Zeng, J., Zhang, D., Lu, D., Mo, P., Li, Z., Chen, Y., Rynik, M., Huang, L., Li, Z., Shi, S., et al. (2023) · The Journal of Chemical Physics · doi:10.1063/5.0155600 · Cited by 652
Ma, W., Cheng, F., Xu, Y., Wen, Q., & Liu, Y. (2019) · Advanced Materials · doi:10.1002/adma.201901111 · Cited by 628
A Critical Review of Machine Learning of Energy Materials
Chen, C., Zuo, Y., Ye, W., Li, X., Deng, Z., & Ong, S. P. (2020) · Advanced Energy Materials · doi:10.1002/aenm.201903242 · Cited by 609
Protein interface prediction using graph convolutional networks
Fout, A., Ben‐Hur, A., Anderson, C., Chitsaz, H., & Zhou, W. (2017) · Digital Collections of Colorado (Colorado State University) · doi:10.25675/3.021826 · Cited by 606
The MLIP package: moment tensor potentials with MPI and active learning
Novikov, I. S., Gubaev, K., Podryabinkin, E. V., & Shapeev, A. V. (2020) · Machine Learning Science and Technology · doi:10.1088/2632-2153/abc9fe · Cited by 631
Large-scale chemical language representations capture molecular structure and properties
Ross, J., Belgodere, B., Chenthamarakshan, V., Padhi, I., Mroueh, Y., & Das, P. (2022) · Nature Machine Intelligence · doi:10.1038/s42256-022-00580-7 · Cited by 590
GaAN: Gated Attention Networks for Learning on Large and Spatiotemporal GraphsPreprint
Zhang, J., Shi, X., Xie, J., Ma, H., King, I., & Yeung, D. (2018) · arXiv (Cornell University) · doi:10.48550/arxiv.1803.07294 · Cited by 636
Machine Learning for Electrocatalyst and Photocatalyst Design and Discovery
Mai, H., Le, T. C., Chen, D., Winkler, D. A., & Caruso, R. A. (2022) · Chemical Reviews · doi:10.1021/acs.chemrev.2c00061 · Cited by 588
Mater, A. C., & Coote, M. L. (2019) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.9b00266 · Cited by 565
Machine-learned potentials for next-generation matter simulations
Friederich, P., Häse, F., Proppe, J., & Aspuru‐Guzik, A. (2021) · Nature Materials · doi:10.1038/s41563-020-0777-6 · Cited by 595
Machine learning molecular dynamics for the simulation of infrared spectra
Gastegger, M., Behler, J., & Marquetand, P. (2017) · Chemical Science · doi:10.1039/c7sc02267k · Cited by 556
Ab initio solution of the many-electron Schrödinger equation with deep neural networks
Pfau, D., Spencer, J. S., Matthews, A. G. D. G., & Foulkes, W. M. C. (2020) · Physical Review Research · doi:10.1103/physrevresearch.2.033429 · Cited by 650
Explainable machine learning in materials science
Zhong, X., Gallagher, B. J., Liu, S., Kailkhura, B., Hiszpanski, A. M., & Han, T. Y.-J. (2022) · npj Computational Materials · doi:10.1038/s41524-022-00884-7 · Cited by 566
Machine Learning of Coarse-Grained Molecular Dynamics Force Fields
Wang, J., Olsson, S., Wehmeyer, C., Pérez, A., Charron, N. E., Fabritiis, G. D., Noé, F., & Clementi, C. (2019) · ACS Central Science · doi:10.1021/acscentsci.8b00913 · Cited by 552
ElemNet: Deep Learning the Chemistry of Materials From Only Elemental Composition
Jha, D., Ward, L. T., Paul, A., Liao, W., Choudhary, A., Wolverton, C. M., & Agrawal, A. (2018) · Scientific Reports · doi:10.1038/s41598-018-35934-y · Cited by 557
Self-Supervised Graph Transformer on Large-Scale Molecular DataPreprint
Rong, Y., Bian, Y., Xu, T., Xie, W., Wei, Y., Huang, W., & Huang, J. (2020) · arXiv (Cornell University) · doi:10.48550/arxiv.2007.02835 · Cited by 552
PotentialNet for Molecular Property Prediction
Feinberg, E. N., Sur, D., Wu, Z., Husic, B. E., Mai, H., Li, Y., Sun, S., Yang, J., Ramsundar, B., & Pande, V. S. (2018) · ACS Central Science · doi:10.1021/acscentsci.8b00507 · Cited by 490
E(n) Equivariant Graph Neural NetworksPreprint
Satorras, V. G., Hoogeboom, E., & Welling, M. (2021) · arXiv (Cornell University) · doi:10.48550/arxiv.2102.09844 · Cited by 481
Machine Learning a General-Purpose Interatomic Potential for Silicon
Bartók, A. P., Kermode, J. R., Bernstein, N., & Cśanyi, G. (2018) · Physical Review X · doi:10.1103/physrevx.8.041048 · Cited by 514
Schütt, K. T., Gastegger, M., Tkatchenko, A., Müller, K. R., & Maurer, R. J. (2019) · Nature Communications · doi:10.1038/s41467-019-12875-2 · Cited by 513
Self-citationThe TensorMol-0.1 model chemistry: a neural network augmented with long-range physics
Yao, K., Herr, J. E., Toth, D. W., Mckintyre, R., & Parkhill, J. (2018) · Chemical Science · doi:10.1039/c7sc04934j · Cited by 489
Convolutional embedding of attributed molecular graphs for physical property prediction
Coley, C. W., Barzilay, R., Green, W. H., Jaakkola, T. S., & Jensen, K. F. (2017) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.6b00601 · Cited by 470
SchNet: A continuous-filter convolutional neural network for modeling quantum interactions
Schütt, K. T., Kindermans, P.-J., Sauceda, H. E., Chmiela, S., Tkatchenko, A., & Müller, K. (2017) · Open Repository and Bibliography (University of Luxembourg) · doi:10.48550/arxiv.1706.08566 · Cited by 463
Self-citationMachine learning for heterogeneous catalyst design and discovery
Goldsmith, B. R., Esterhuizen, J. A., Liu, J., Bartel, C. J., & Sutton, C. A. (2018) · AIChE Journal · doi:10.1002/aic.16198 · Cited by 460
The Role of Machine Learning in the Understanding and Design of Materials
Moosavi, S. M., Jablonka, K. M., & Smit, B. (2020) · Journal of the American Chemical Society · doi:10.1021/jacs.0c09105 · Cited by 451
SchNetPack: A Deep Learning Toolbox For Atomistic Systems
Schütt, K. T., Kessel, P., Gastegger, M., Nicoli, K. A., Tkatchenko, A., & Müller, K. R. (2018) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.8b00908 · Cited by 453
Self-citationMachine Learning for Electronically Excited States of Molecules
Westermayr, J., & Marquetand, P. (2020) · Chemical Reviews · doi:10.1021/acs.chemrev.0c00749 · Cited by 445
Quantum Chemistry in the Age of Machine Learning
Dral, P. O. (2020) · The Journal of Physical Chemistry Letters · doi:10.1021/acs.jpclett.9b03664 · Cited by 436
Graph convolutional networks for computational drug development and discovery
Sun, M., Zhao, S., Gilvary, C. M., Elemento, O., Zhou, J., & Wang, F. (2019) · Briefings in Bioinformatics · doi:10.1093/bib/bbz042 · Cited by 424
Unsupervised Learning Methods for Molecular Simulation Data
Glielmo, A., Husic, B. E., Rodríguez, Á., Clementi, C., Noé, F., & Laio, A. (2021) · Chemical Reviews · doi:10.1021/acs.chemrev.0c01195 · Cited by 436
Lim, J., Ryu, S., Park, K., Choe, Y. J., Ham, J., & Kim, W. Y. (2019) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.9b00387 · Cited by 411
A cloud detection algorithm for satellite imagery based on deep learning
Jeppesen, J. H., Jacobsen, R. H., Inceoglu, F., & Toftegaard, T. S. (2019) · Remote Sensing of Environment · doi:10.1016/j.rse.2019.03.039 · Cited by 402
Deep learning in bioinformatics: Introduction, application, and perspective in the big data era
Li, Y., Huang, C., Ding, L., Li, Z., Pan, Y., & Gao, X. (2019) · Methods · doi:10.1016/j.ymeth.2019.04.008 · Cited by 401
Time-lagged autoencoders: Deep learning of slow collective variables for molecular kinetics
Wehmeyer, C., & Noé, F. (2018) · The Journal of Chemical Physics · doi:10.1063/1.5011399 · Cited by 422
Artificial intelligence for geoscience: Progress, challenges, and perspectives
Zhao, T., Wang, S., Ouyang, C., Chen, M., Liu, C., Zhang, J., Long, Y., Wang, F., Xie, Y., Li, J., et al. (2024) · The Innovation · doi:10.1016/j.xinn.2024.100691 · Cited by 409
Unified representation of molecules and crystals for machine learning
Huo, H., & Rupp, M. (2022) · Machine Learning Science and Technology · doi:10.1088/2632-2153/aca005 · Cited by 419
Quantum chemical accuracy from density functional approximations via machine learning
Bogojeski, M., Vogt-Maranto, L., Tuckerman, M. E., Müller, K., & Burke, K. (2020) · Nature Communications · doi:10.1038/s41467-020-19093-1 · Cited by 401
Self-citationDeep learning model to predict complex stress and strain fields in hierarchical composites
Yang, Z., Yu, C., & Buehler, M. J. (2021) · Science Advances · doi:10.1126/sciadv.abd7416 · Cited by 399
Geometric deep learning on molecular representations
Atz, K., Grisoni, F., & Schneider, G. (2021) · Nature Machine Intelligence · doi:10.1038/s42256-021-00418-8 · Cited by 415
Exploring chemical compound space with quantum-based machine learning
von Lilienfeld, O. A., Müller, K., & Tkatchenko, A. (2020) · Nature Reviews Chemistry · doi:10.1038/s41570-020-0189-9 · Cited by 396
Self-citationInsightful classification of crystal structures using deep learning
Ziletti, A., Kumar, D., Scheffler, M., & Ghiringhelli, L. M. (2018) · Nature Communications · doi:10.1038/s41467-018-05169-6 · Cited by 383
Machine learning: Accelerating materials development for energy storage and conversion
Chen, A., Zhang, X., & Zhou, Z. P. (2020) · InfoMat · doi:10.1002/inf2.12094 · Cited by 381
Lagrangian Neural NetworksPreprint
Cranmer, M., Greydanus, S., Hoyer, S., Battaglia, P., Spergel, D. N., & Ho, S. (2020) · arXiv (Cornell University) · doi:10.48550/arxiv.2003.04630 · Cited by 611
Autonomous Discovery in the Chemical Sciences Part I: Progress
Coley, C. W., Eyke, N. S., & Jensen, K. F. (2019) · Angewandte Chemie International Edition · doi:10.1002/anie.201909987 · Cited by 371
Alchemical and structural distribution based representation for universal quantum machine learning
Faber, F. A., Christensen, A. S., Huang, B., & von Lilienfeld, O. A. (2018) · The Journal of Chemical Physics · doi:10.1063/1.5020710 · Cited by 372
Machine learning for interatomic potential models
Mueller, T., Hernández, A., & Wang, C. (2020) · The Journal of Chemical Physics · doi:10.1063/1.5126336 · Cited by 380
Metadata
- Authors
- Schütt, K. T., Arbabzadah, F., Chmiela, S., Müller, K. R., & Tkatchenko, A.
- Affiliations
- Technische Universität Berlin (DE)
- Korea University (KR)
- University of Luxembourg (LU)
- Corresponding author
- Müller, Klaus R.
- Tkatchenko, Alexandre
- Venue
- Nature Communications
- Published
- 2017-01-09
- Volume
- 8
- Issue
- 1
- Pages
- 13890-13890
- ISSN
- 2041-1723 (ISSN-L)
- PMC
- PMC5228054
- OpenAlex
- W2527189750
- Cited by
- 1,445
- References
- 55
- Citations by year
- 2026: 89 · 2025: 155 · 2024: 167 · 2023: 170 · 2022: 147 · 2021: 182 · 2020: 213 · 2019: 164 · 2018: 126 · 2017: 29
- FWCI
- 61.66
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Locations
- Nature Communications (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- arXiv (Cornell University) (submittedVersion)
- MPG.PuRe (Max Planck Society) (submittedVersion, cc-by)
- RePEc: Research Papers in Economics (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- Open Repository and Bibliography (University of Luxembourg) (submittedVersion, cc-by-nc-sa)
- arXiv (Cornell University)
- Funders
- National Science Foundation
- National Research Foundation (2012-005741)
- Deutsche Forschungsgemeinschaft (MU 987/20-1, MU 987/20)
- Bundesministerium für Bildung und Forschung (01 IS 14013 A)
- Banting and Best Diabetes Centre, University of Toronto (01IS14013A)
- Topics
- Physical Sciences › Materials Science › Materials Chemistry › Machine Learning in Materials Science
- Physical Sciences › Physics and Astronomy › Atomic and Molecular Physics, and Optics › Advanced Chemical Physics Studies
- Life Sciences › Pharmacology, Toxicology and Pharmaceutics › Drug Discovery › Computational Drug Discovery Methods
- Keywords
- many-body Hamiltonian, molecular energy prediction, local chemical potential, atomic energy, electronic structure, quantum many-body systems
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
