

The DOI matches its Crossref record; 37 of the 37 references it cites were found in a registry or an index.
GC-Content Normalization for RNA-Seq Data
Open accessAbstract
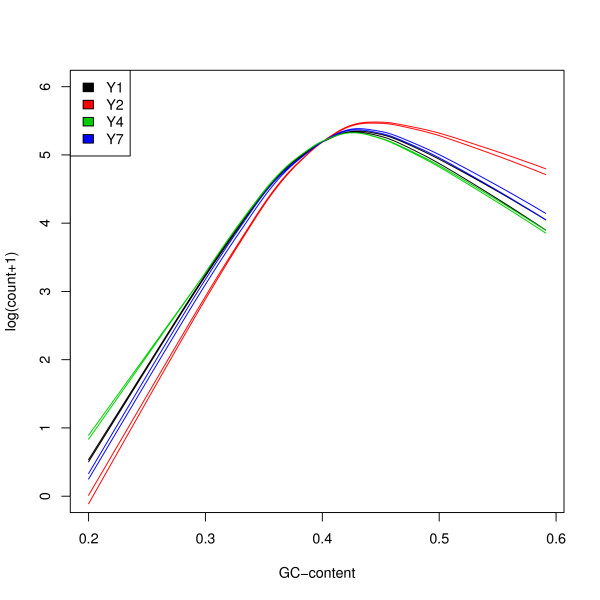
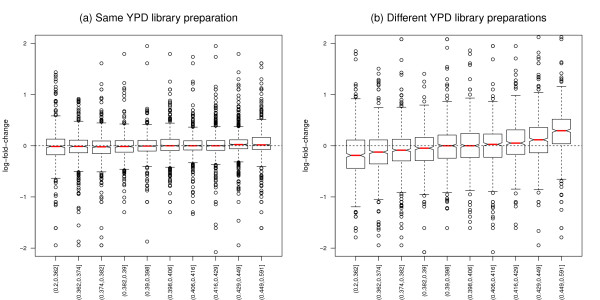
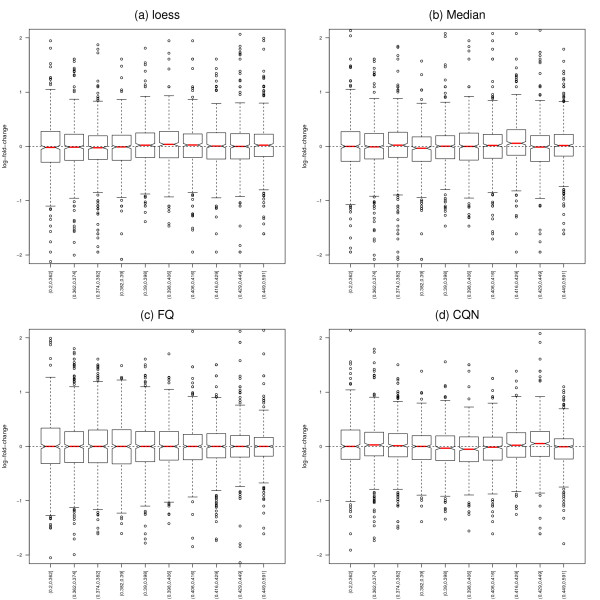
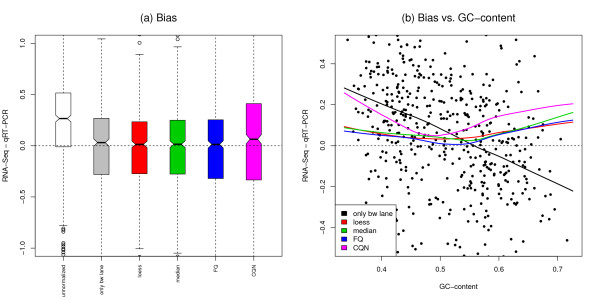
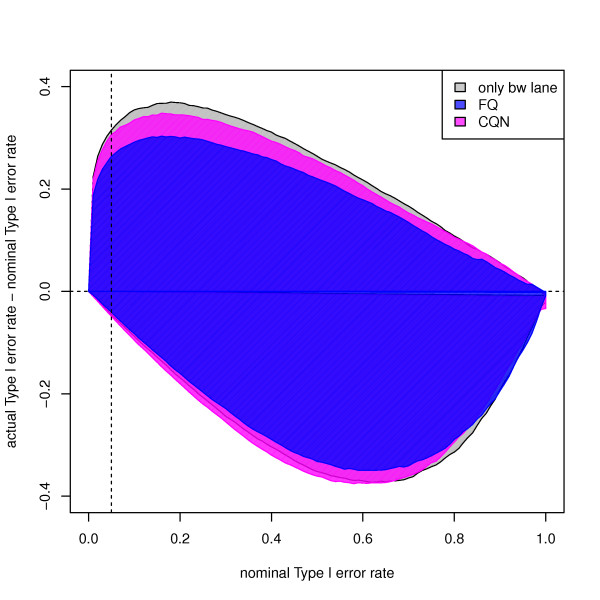
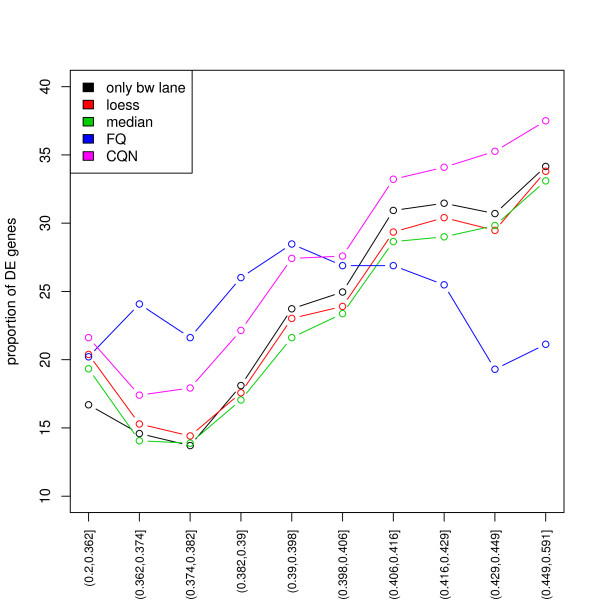
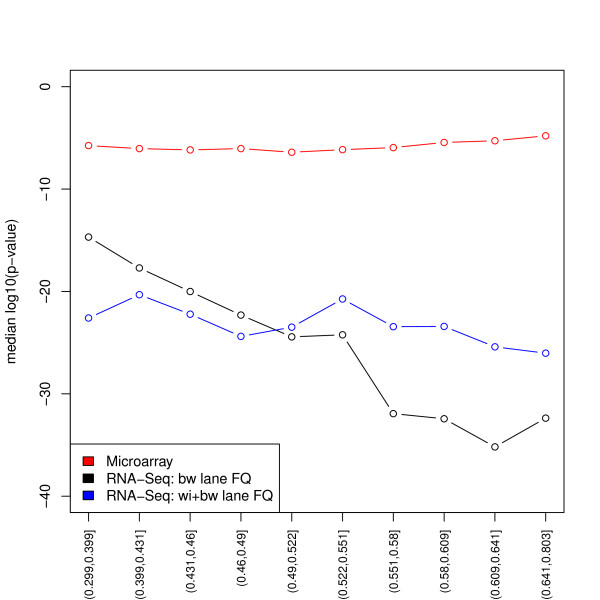
Background: Transcriptome sequencing (RNA-Seq) has become the assay of choice for high-throughput studies of gene expression. However, as is the case with microarrays, major technology-related artifacts and biases affect the resulting expression measures. Normalization is therefore essential to ensure accurate inference of expression levels and subsequent analyses thereof. Results: We focus on biases related to GC-content and demonstrate the existence of strong sample-specific GC-content effects on RNA-Seq read counts, which can substantially bias differential expression analysis. We propose three simple within-lane gene-level GC-content normalization approaches and assess their performance on two different RNA-Seq datasets, involving different species and experimental designs. Our methods are compared to state-of-the-art normalization procedures in terms of bias and mean squared error for expression fold-change estimation and in terms of Type I error and p-value distributions for tests of differential expression. The exploratory data analysis and normalization methods proposed in this article are implemented in the open-source Bioconductor R package EDASeq. Conclusions: Our within-lane normalization procedures, followed by between-lane normalization, reduce GC-content bias and lead to more accurate estimates of expression fold-changes and tests of differential expression. Such results are crucial for the biological interpretation of RNA-Seq experiments, where downstream analyses can be sensitive to the supplied lists of genes.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“commonly used methods fornormalization (cqn, EDASeq, RUV2, sva, PEER) varied in their ability to remove these systematic biases, depending on sample complexity and initial data quality.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
Experimental — accuracy not yet measuredIn the citing paper
“These effects can be minimized by appropriate experimental design [51] or, alternatively, removed by batch-correction methods such as COMBAT [52] or ARSyN [20, 53].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“Filtering out genes that are expressed at low levels prior to differential expression analysis reduces the severity of the correction and may improve the power of detection [20].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“NOISeq and NOISeqBIO (the adaptation of NOISeq for biological replication) are more efficient in avoiding false positive calls at the cost of some sensitivity but perform well with different numbers of replicates [10, 20, 212].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“SAMseq performs well in terms of FDR but presents an acceptable sensitivity when the number of replicates is relatively high, at least 10 [20, 55, 209].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“Notably, neither applying GC bias correction tools (cqn14 and EDASeq15) to individual sites nor changing read counts to only use 3′-UTRs was effective at decreasing the number of inter-site false positive DEGs (Fig. 1b and Supplementary Fig. 3); in most cases, these methods actually increased the number of false positives.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
In the citing paper
“The application of GC content bias correction packages including cqn14 and EDASeq15 could not remove these false positives, likely because GC content bias is not the only source that contributes to bias in gene expression data.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
In the citing paper
“This confirms the impact of two already recognized RNA-seq latent variables, GC-content and insert size15,22, and it also identifies two more relevant contributions to technical variation, gene coverage variation and error rate.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Controlling the False Discovery Rate: A Practical and Powerful Approach to Multiple Testing
Benjamini, Y., & Hochberg, Y. (1995) · Journal of the Royal Statistical Society Series B: Statistical Methodology · doi:10.1111/j.2517-6161.1995.tb02031.x · Cited by 109,482
Robinson, M. D., McCarthy, D. J., & Smyth, G. K. (2010) · Bioinformatics · doi:10.1093/bioinformatics/btp616 · Cited by 41,364
Ultrafast and memory-efficient alignment of short DNA sequences to the human genome.
Langmead, B., Trapnell, C., Pop, M., & Salzberg, S. L. (2009) · Genome Biology · doi:10.1186/gb-2009-10-3-r25 · Cited by 21,849
Differential expression analysis for sequence count data
Anders, S., & Huber, W. (2010) · Genome Biology · doi:10.1186/gb-2010-11-10-r106 · Cited by 15,581
Mapping and quantifying mammalian transcriptomes by RNA-Seq
Mortazavi, A., Williams, B. A., McCue, K., Schaeffer, L., & Wold, B. (2008) · Nature Methods · doi:10.1038/nmeth.1226 · Cited by 13,696
RNA-Seq: a revolutionary tool for transcriptomics
Wang, Z., Gerstein, M., & Snyder, M. (2009) · Nature Reviews Genetics · doi:10.1038/nrg2484 · Cited by 13,174
Smyth, G. K. (2004) · Statistical Applications in Genetics and Molecular Biology · doi:10.2202/1544-6115.1027 · Cited by 11,829
Exploration, normalization, and summaries of high density oligonucleotide array probe level data
Irizarry, R. A. (2003) · Biostatistics · doi:10.1093/biostatistics/4.2.249 · Cited by 10,603
A scaling normalization method for differential expression analysis of RNA-seq data
Robinson, M. D., & Oshlack, A. (2010) · Genome Biology · doi:10.1186/gb-2010-11-3-r25 · Cited by 7,876
Gene ontology analysis for RNA-seq: accounting for selection bias
Young, M. D., Wakefield, M. J., Smyth, G. K., & Oshlack, A. (2010) · Genome Biology · doi:10.1186/gb-2010-11-2-r14 · Cited by 7,107
Cited by
Sorted by citations
Items 701–800 of 865
Functional Interpretation of High-Throughput Sequencing Data.
Lee, C. K. (2016) · Deep Blue (University of Michigan)
Sedaghat, N., Stanway, I. B., Zangeneh, S. Z., Bammler, T. K., & Shojaie, A. (2017) · Elsevier eBooks · doi:10.1016/b978-0-12-801238-3.64209-5
Samra, S., Tokovenko, B. T., Kaszubowska, M., Makitrynskyy, R., Alali, A., Schmidt, Y., Welle, E., Gross, S., Paululat, T., Luzhetskyy, A. N., et al. (2018) · Natural Products Chemistry & Research · doi:10.4172/2329-6836.1000333
Nucleic Acids: RNA Identification and Quantification Via Next-Generation Sequencing
Buschmann, D., Kirchner, B., Mussack, V., & Pfaffl, M. W. (2018) · Elsevier eBooks · doi:10.1016/b978-0-12-409547-2.14269-6
Sabedot, T. S. (2019) · Digital Library of Theses and Dissertations (Universidade de São Paulo) · doi:10.11606/t.17.2019.tde-17102018-142049
The Role of Exploratory Data Analysis and Pre-processing in Omics Studies
Schiffman, C. (2019) · eScholarship (California Digital Library)
Chassé, M. H., Johnson, B. K., Boguslawski, E. A., Sorensen, K. M., Heo, L., Madaj, Z., Beddows, I., Foxa, G. E., Kitchen-Goosen, S. M., Williams, B. O., et al. (2019) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/861484
Transcriptome-based identification of small RNA in plants: The need for robust prediction algorithms
Ganguli, S., Singh, P. K., & Pal, A. (2020) · Elsevier eBooks · doi:10.1016/b978-0-12-817112-7.00004-3
A Quantitative Comparisons of $\beta$-Coronavirus Genomes and Their Associated GenesPreprint
Hassan, S. S., & Rout, R. K. (2020) · Preprints.org · doi:10.20944/preprints202004.0486.v1
Genetic variation as a tool for identifying novel transducers of itch
Morita, T. (2016) · eScholarship (California Digital Library)
Time-Course Analysis and Clustering of Gene Expression Data
DeGraaf, S. (2020) · eScholarship (California Digital Library)
An IPI based immune prognostic model for diffuse large B-cell lymphomaPreprint
Mu, S., Shi, D., Ai, L., Fan, F., Peng, F., Sun, C., & Hu, Y. (2021) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2021.03.03.433839
Lobo, D., Godinho, R., & Archer, J. P. (2021) · Research Square · doi:10.21203/rs.3.rs-468506/v1
RNA sequencing of non-coding RNAs in ischaemic heart disease
Ward, Z. (2021) · Otago University Research Archive (University of Otago)
Taming of the wild: a new method for cross study RNA-seq analysisPreprint
Lobo, D., Godinho, R., & Archer, J. P. (2020) · doi:10.21203/rs.3.rs-27674/v1
Construction of transcriptional regulatory networks using total RNA-seq data
Chouvarine, P., & Hansmann, G. (2021) · STAR Protocols · doi:10.1016/j.xpro.2021.100769
Unveiling the two faces of immune dysregulation in Systemic Sclerosis
Servaas, N. H. (2021) · Utrecht University Repository (Utrecht University) · doi:10.33540/93
Williamson, V. S. (2012) · VCU Scholars Compass (Virginia Commonwealth University) · doi:10.25772/seyx-fq65
Vu, N., & Uyen, B. U. I. (2021) · MOL2NET International Conference on Multidisciplinary Sciences (MOL2NET) · doi:10.3390/mol2net-07-11601
Angers, S., & MacLeod, G. D. A. H. (2021) · Faculty Opinions – Post-Publication Peer Review of the Biomedical Literature · doi:10.3410/f.739341152.793585340
(2016) · CRC Press eBooks · doi:10.1201/b19532-22
Hotz, H.-R. (2019) · Faculty Opinions – Post-Publication Peer Review of the Biomedical Literature · doi:10.3410/f.736736304.793566365
Joseph, P. (2022) · Wiley eBooks · doi:10.1002/9781119807704.ch7
Troeberg, L. (2019) · Faculty Opinions – Post-Publication Peer Review of the Biomedical Literature · doi:10.3410/f.735001574.793559769
journal, E. (2012) · EMBnet journal · doi:10.14806/ej.18.a.497
journal, E. (2012) · EMBnet journal · doi:10.14806/ej.18.b.592
Ried, T., & Hirsch, D. D. (2018) · Faculty Opinions – Post-Publication Peer Review of the Biomedical Literature · doi:10.3410/f.732011041.793545090
Evans, D. L. (2021) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.22541/au.162740437.78291655/v1
Bahram, S., & Follain, G. (2018) · Faculty Opinions – Post-Publication Peer Review of the Biomedical Literature · doi:10.3410/f.734001323.793552363
Polysomal mRNA Association and Gene Expression in Trypanosoma brucei
Tinti, M., Kelner-Mirôn, A., Marriott, L. J., & Ferguson, M. A. J. (2021) · Wellcome Open Research · doi:10.12688/wellcomeopenres.16430.2
Transcriptome Analysis Using RNA Sequencing for Finding Genes Related to Fiber in Cotton: A Review
Etukuri, S. P., Anche, V. C., Sobitjonovich, A. M., Walker, L. T., & Sripathi, V. R. (2022) · IntechOpen eBooks · doi:10.5772/intechopen.104572
Lutter, L. (2022) · Utrecht University Repository (Utrecht University) · doi:10.33540/1160
Genomic and Proteomic Analysis of Patients with Reoccurring Discrete Subaortic StenosisPreprint
Birla, R. K., Singampalli, K. L., Nieuwsma, A., Brimmer, S., Kaul, A., Wadhwa, L., Jung, S. Y., Mezquita, D., Grimm, S. L., Balaji, S., et al. (2023) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2023.01.22.525022
Intra-Exon Motif Correlations as a Proxy Measure for Mean Per-Tile Sequence Quality Data in RNA-Seq
Alnasir, J. J., & Shanahan, H. P. (2023) · Journal of Computational Biology · doi:10.1089/cmb.2021.0476
Sarmah, D. T., Kumar, S., Chatterjee, S., & Bairagi, N. (2023) · Elsevier eBooks · doi:10.1016/b978-0-323-85713-0.00015-3
Emrich, S. M., Yoast, R. E., Zhang, X., Fike, A. J., Wang, Y., Bricker, K. N., Tao, A. Y., Xin, P., Walter, V., Johnson, M. T., et al. (2023) · eLife · doi:10.7554/elife.84708.sa2
Rodrigues, T. S. (2022) · Digital Library of Theses and Dissertations (Universidade de São Paulo) · doi:10.11606/t.17.2022.tde-10042023-161658
Turan, T., Kongpachith, S., Halliwill, K. D., McLaughlin, R. T., Binnewies, M., Reddy, D., Zhao, X., Mathew, R. S., Ye, S., Jacob, H. J., et al. (2023) · doi:10.1158/2326-6066.22812519
Turan, T., Kongpachith, S., Halliwill, K. D., McLaughlin, R. T., Binnewies, M., Reddy, D., Zhao, X., Mathew, R. S., Ye, S., Jacob, H. J., et al. (2023) · doi:10.1158/2326-6066.23283442
Lambrechts, Y., Hatse, S., Richard, F., Boeckx, B., Floris, G., Desmedt, C., Smeets, A., Neven, P., Lambrechts, D., & Wildiers, H. P. M. W. (2023) · Cancers · doi:10.3390/cancers15174341
Li, T., Tang, X., & Chatrath, A. (2023) · The Annals of Applied Statistics · doi:10.1214/23-aoas1771
Functional characterization of long non-coding RNAs in Schistosoma mansoni
Silveira, G. O. (2022) · Digital Library of Theses and Dissertations (Universidade de São Paulo) · doi:10.11606/t.46.2022.tde-07112023-145651
Ricci, R., Fletcher, J. L., Makowiecki, K., Pepper, R., Fortune, A. J., Cullen, C. L., Connelly, W. M., Charlesworth, J., Blackburn, N. B., Pitman, K. A., et al. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.02.29.582867
Robust estimation of cancer and immune cell-type proportions from bulk tumor ATAC-Seq dataPreprint
Gabriel, A. A., Racle, J., Falquet, M., Jandus, C., & Gfeller, D. (2024) · eLife · doi:10.7554/elife.94833.1
Dong, K., Chen, W., Pan, X., Wang, H., Sun, Y., Cheng, Q., Chen, W., Wang, C., Fu, Y., & Cui, X. (2022) · Figshare · doi:10.6084/m9.figshare.19126403
Additional file 8 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424934
Dong, K., Chen, W., Pan, X., Wang, H., Sun, Y., Cheng, Q., Chen, W., Wang, C., Fu, Y., & Cui, X. (2022) · Figshare · doi:10.6084/m9.figshare.19126397
Loubet, I., Meyer, L. S., Michel, S., Pernin, F., Carrère, S., Barrès, B., Corre, V. L., & Délye, C. (2023) · Figshare · doi:10.6084/m9.figshare.24432033
Dong, K., Chen, W., Pan, X., Wang, H., Sun, Y., Cheng, Q., Chen, W., Wang, C., Fu, Y., & Cui, X. (2022) · Figshare · doi:10.6084/m9.figshare.19126400
Additional file 9 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424937
Dong, K., Chen, W., Pan, X., Wang, H., Sun, Y., Cheng, Q., Chen, W., Wang, C., Fu, Y., & Cui, X. (2022) · Figshare · doi:10.6084/m9.figshare.19126406
Statistical methods for the analysis of complex epidemiology data
Acebes, M. E. (2023) · UPCommons institutional repository (Universitat Politècnica de Catalunya) · doi:10.5821/dissertation-2117-407878
Genomic Feature of Filled Regions of Telomere-to-Telomere Genomes in Five Model SpeciesPreprint
Xiong, C., Zhang, H., Zhang, Y., Hao, L., Liang, Z., & Zheng, J. (2024) · SSRN Electronic Journal · doi:10.2139/ssrn.4853623
Robust estimation of cancer and immune cell-type proportions from bulk tumor ATAC-Seq dataPreprint
Gabriel, A. A., Racle, J., Falquet, M., Jandus, C., & Gfeller, D. (2024) · eLife · doi:10.7554/elife.94833.2
Robust estimation of cancer and immune cell-type proportions from bulk tumor ATAC-Seq dataPreprint
Gabriel, A. A., Racle, J., Falquet, M., Jandus, C., & Gfeller, D. (2024) · eLife · doi:10.7554/elife.94833.3
Gabriel, A. A., Racle, J., Falquet, M., Jandus, C., & Gfeller, D. (2024) · eLife · doi:10.7554/elife.94833.4.sa3
Integrative Bioinformatics: Databases, Web Portals, and Tools
Settino, M., & Cannataro, M. (2024) · Elsevier eBooks · doi:10.1016/b978-0-323-95502-7.00132-9
Benefits and challenges of OMICS data integration at the pathway level
Borisov, N. M., & Sorokin, M. (2024) · Elsevier eBooks · doi:10.1016/b978-0-443-15568-0.00010-8
High-Accuracy RNA Integrity Definition for Unbiased Transcriptome Comparisons with INDEGRAPreprint
Cleynen, A., Ravindran, A., Sethi, A. J., Kumar, B. R. P., Javaid, T., Mahmud, S., Woodward, K., Vieira, H. G. S., Änkö, M., Weatheritt, R. J., et al. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.12.12.627949
SFSWAP is a negative regulator of OGT intron detention and global pre-mRNA splicingPreprint
Govindan, A., & Conrad, N. K. (2024) · eLife · doi:10.7554/elife.104439.1
Beyond the network-based multi-omics data integration in COPD: a pathway-centric analysis
Sibilio, P., Conte, F., & Paci, P. (2024) · IEEE International Conference on Bioinformatics and Biomedicine (BIBM) · doi:10.1109/bibm62325.2024.10822251
Roggisch, J. (2025) · SSRN Electronic Journal · doi:10.2139/ssrn.5170682
SFSWAP is a negative regulator of OGT intron detention and global pre-mRNA splicingPreprint
Govindan, A., & Conrad, N. K. (2025) · eLife · doi:10.7554/elife.104439.2
Author response: SFSWAP is a negative regulator of OGT intron detention and global pre-mRNA splicing
Govindan, A., & Conrad, N. K. (2025) · eLife · doi:10.7554/elife.104439.3.sa4
Jabeen, A., Awartani, D., Sherif, S., Ahmed, E. I., Alanany, R., Saleh, A., Hendrickx, W., & Raynaud, C. M. (2026) · British Journal of Cancer · doi:10.1038/s41416-026-03571-2
Hulbert, A., Tiv, R., Cao, B. L., Cesario, D., Mallisetty, A., Gauchat, E., Rana, A., Weinberg, F. D., Danilova, L., & Jusué-Torres, I. (2025) · Journal of Thoracic Oncology · doi:10.1016/j.jtho.2025.08.015
Sedaghat, N., Stanway, I. B., Zangeneh, S. Z., Bammler, T. K., & Shojaie, A. (2018) · Elsevier eBooks · doi:10.1016/b978-0-323-95488-4.10022-1
Concomitant acetylation and loading of H2A.Z by NuA4/TIP60 regulate target gene transcriptionPreprint
Mameri, A., Lachance, C., Cattoglio, C., Bianco, S., Humbert, J., Joly-Beauparlant, C., Texeraud, E., Banerjea, A., Lashgari, A., Lambert, J., et al. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.09.22.677705
Contrasting roles of KMT2C and KMT2D in breast cancer
Tinsley, E. A., Bredin, P., Toomey, S., Hennessy, B. T., & Furney, S. J. (2025) · Scientific Reports · doi:10.1038/s41598-025-19027-1
Kelley, L. H., Houlihan, A. S., Yongblah, K., Aljiboury, A. A., Davis, S. L., & Maine, E. M. (2025) · Developmental Biology · doi:10.1016/j.ydbio.2025.10.013
Abundant Parent-of-origin Effect eQTL: The Framingham Heart StudyPreprint
Guan, Y., Huan, T., & Levy, D. A. (2025) · eLife · doi:10.7554/elife.107920
Author response: Abundant Parent-of-origin Effect eQTL: The Framingham Heart Study
Guan, Y., Huan, T., & Levy, D. A. (2025) · eLife · doi:10.7554/elife.107920.1.sa0
Abundant Parent-of-origin Effect eQTL: The Framingham Heart StudyPreprint
Guan, Y., Huan, T., & Levy, D. A. (2025) · eLife · doi:10.7554/elife.107920.1
Osabe, T., Shimizu, K., & Kadota, K. (2021) · Figshare · doi:10.6084/m9.figshare.16841049
Osabe, T., Shimizu, K., & Kadota, K. (2021) · Figshare · doi:10.6084/m9.figshare.16841061
Osabe, T., Shimizu, K., & Kadota, K. (2021) · Figshare · doi:10.6084/m9.figshare.16841058
Almeida, S. F. F., Santos, L., Sampaio-Ribeiro, G., Ferreira, H. R. S., Lima, N. R., Caetano, R., Abreu, M., Zuzarte, M., Ribeiro, A. S., Paiva, A., et al. (2023) · Figshare · doi:10.6084/m9.figshare.24669831
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716600
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716627
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716606
Kakati, T., Bhattacharyya, D. K., Kalita, J. K., & Norden‐Krichmar, T. M. (2022) · Figshare · doi:10.6084/m9.figshare.17982050
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716594
Simeone, I., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26564907
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716603
Liang, L., He, M., Zhang, Y., Wang, C., Qin, Z., Li, Q., Yang, T., Meng, F., Zhou, Y., Ge, H., et al. (2023) · Figshare · doi:10.6084/m9.figshare.24709051
Additional file 7 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424931
Dittlau, K. S., Terrie, L., Baatsen, P., Kerstens, A., Swert, L. D., Janky, R., Corthout, N., Masrori, P., Damme, P. V., Hyttel, P., et al. (2024) · Figshare · doi:10.6084/m9.figshare.26562262
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716621
Additional file 1 of The CBP KIX domain regulates long-term memory and circadian activity
Chatterjee, S., Angelakos, C. C., Bahl, E., Hawk, J. D., Gaine, M. E., Poplawski, S. G., Schneider‐Anthony, A., Yadav, M., Porcari, G. S., Cassel, J., et al. (2020) · Figshare · doi:10.6084/m9.figshare.13167354
Additional file 5 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424925
Liang, L., He, M., Zhang, Y., Wang, C., Qin, Z., Li, Q., Yang, T., Meng, F., Zhou, Y., Ge, H., et al. (2024) · Figshare · doi:10.6084/m9.figshare.26647374
Loubet, I., Meyer, L. S., Michel, S., Pernin, F., Carrère, S., Barrès, B., Corre, V. L., & Délye, C. (2024) · Figshare · doi:10.6084/m9.figshare.24432085
Caruso, F. P., D'Andrea, M. R., Coppola, L., Landriscina, M., Condelli, V., Cerulo, L., Giordano, G., Porrás, A., & Pancione, M. (2022) · Figshare · doi:10.6084/m9.figshare.20479635
Costa, M. C., Angelini, C., Franzese, M., Iside, C., Salvatore, M., Laezza, L., Napolitano, F., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26693919
Metadata
- Authors
- Risso, D., Schwartz, K., Sherlock, G., & Dudoit, S.
- Affiliations
- University of Padua (IT)
- Stanford University (US)
- University of California, Berkeley (US)
- Corresponding author
- Dudoit, Sandrine
- Venue
- BMC Bioinformatics
- Published
- 2011-12
- Volume
- 12
- Issue
- 1
- Pages
- 480-480
- ISSN
- 1471-2105 (ISSN-L)
- PMC
- PMC3315510
- OpenAlex
- W2168342464
- Cited by
- 867
- References
- 35
- Citations by year
- 2026: 29 · 2025: 58 · 2024: 83 · 2023: 80 · 2022: 89 · 2021: 97 · 2020: 74 · 2019: 55 · 2018: 60 · 2017: 49 · 2016: 59 · 2015: 38 · 2014: 48 · 2013: 30 · 2012: 18
- FWCI
- 12.48
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- cc-by
- Locations
- BMC Bioinformatics (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Collection of Biostatistics Research Archive (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- eScholarship (California Digital Library) (submittedVersion)
- Padua Research Archive (University of Padova) (submittedVersion, cc-by)
- Funders
- Università degli Studi di Padova
- National Institutes of Health
- National Human Genome Research Institute (R01 HG03468, R01HG003468)
- National Institute of Allergy and Infectious Diseases (R01AI077737)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene expression and cancer classification
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Keywords
- RNA-seq, differential gene expression, gene expression bias, transcriptome sequencing
- MeSH terms
- Base Composition, Saccharomyces cerevisiae, Sequence Analysis, RNA, Gene Expression Profiling, Transcriptome
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
