

The DOI matches its Crossref record; 37 of the 37 references it cites were found in a registry or an index.
GC-Content Normalization for RNA-Seq Data
Open accessAbstract
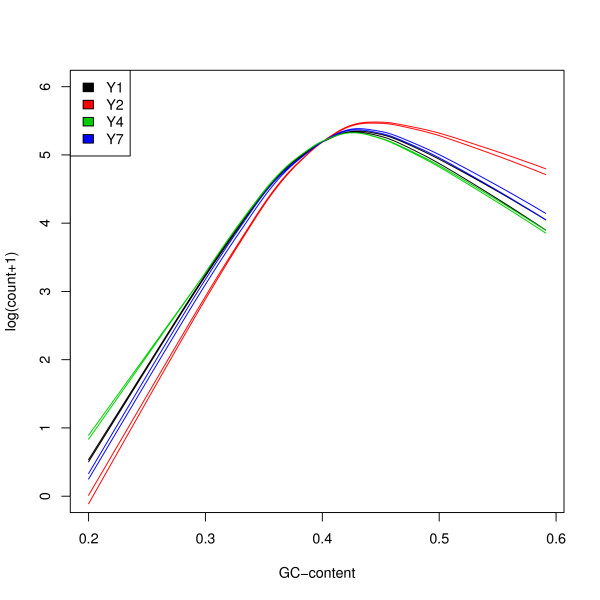
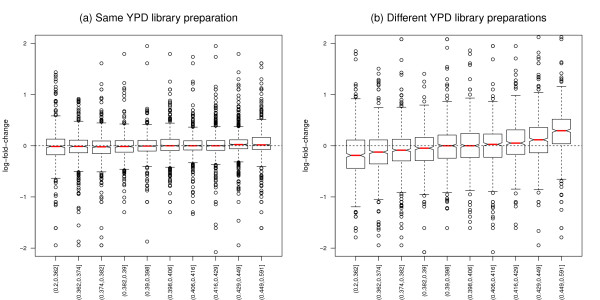
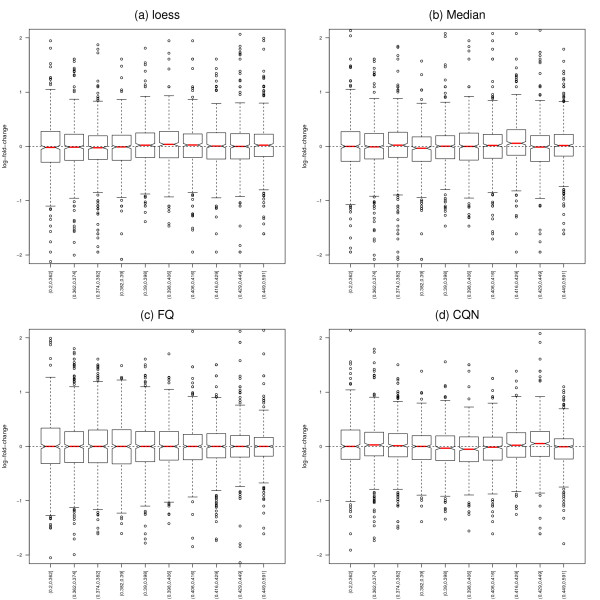
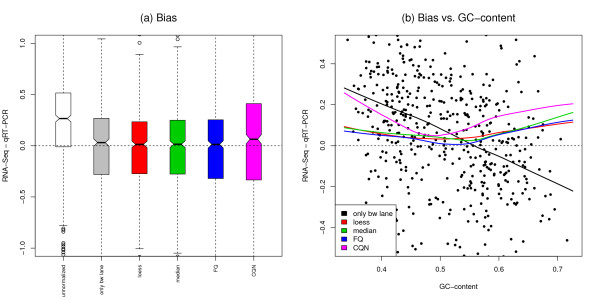
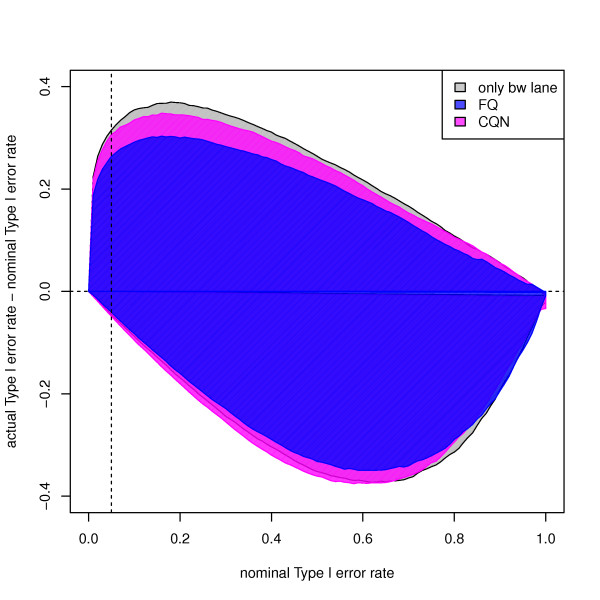
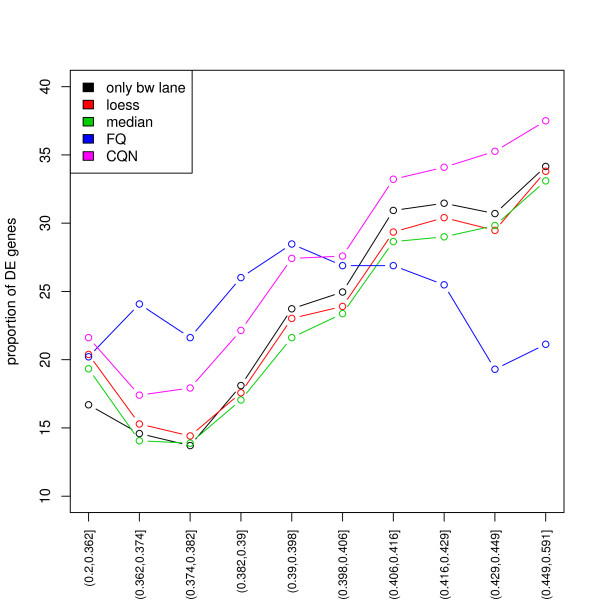
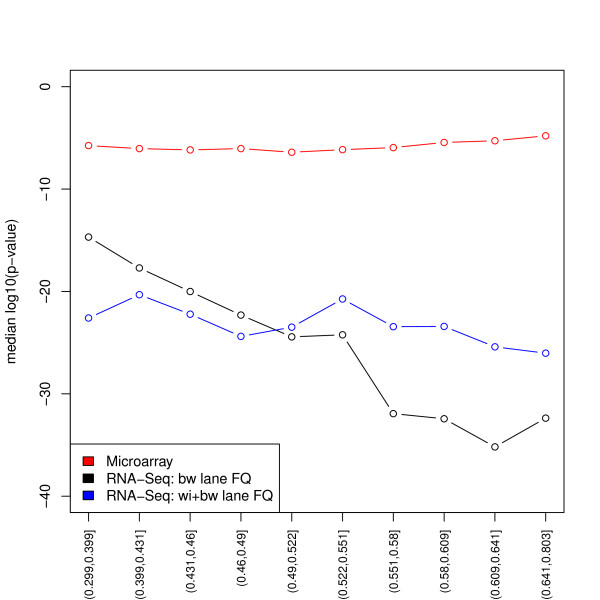
Background: Transcriptome sequencing (RNA-Seq) has become the assay of choice for high-throughput studies of gene expression. However, as is the case with microarrays, major technology-related artifacts and biases affect the resulting expression measures. Normalization is therefore essential to ensure accurate inference of expression levels and subsequent analyses thereof. Results: We focus on biases related to GC-content and demonstrate the existence of strong sample-specific GC-content effects on RNA-Seq read counts, which can substantially bias differential expression analysis. We propose three simple within-lane gene-level GC-content normalization approaches and assess their performance on two different RNA-Seq datasets, involving different species and experimental designs. Our methods are compared to state-of-the-art normalization procedures in terms of bias and mean squared error for expression fold-change estimation and in terms of Type I error and p-value distributions for tests of differential expression. The exploratory data analysis and normalization methods proposed in this article are implemented in the open-source Bioconductor R package EDASeq. Conclusions: Our within-lane normalization procedures, followed by between-lane normalization, reduce GC-content bias and lead to more accurate estimates of expression fold-changes and tests of differential expression. Such results are crucial for the biological interpretation of RNA-Seq experiments, where downstream analyses can be sensitive to the supplied lists of genes.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“commonly used methods fornormalization (cqn, EDASeq, RUV2, sva, PEER) varied in their ability to remove these systematic biases, depending on sample complexity and initial data quality.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
Experimental — accuracy not yet measuredIn the citing paper
“These effects can be minimized by appropriate experimental design [51] or, alternatively, removed by batch-correction methods such as COMBAT [52] or ARSyN [20, 53].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“Filtering out genes that are expressed at low levels prior to differential expression analysis reduces the severity of the correction and may improve the power of detection [20].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“NOISeq and NOISeqBIO (the adaptation of NOISeq for biological replication) are more efficient in avoiding false positive calls at the cost of some sensitivity but perform well with different numbers of replicates [10, 20, 212].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“SAMseq performs well in terms of FDR but presents an acceptable sensitivity when the number of replicates is relatively high, at least 10 [20, 55, 209].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“Notably, neither applying GC bias correction tools (cqn14 and EDASeq15) to individual sites nor changing read counts to only use 3′-UTRs was effective at decreasing the number of inter-site false positive DEGs (Fig. 1b and Supplementary Fig. 3); in most cases, these methods actually increased the number of false positives.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
In the citing paper
“The application of GC content bias correction packages including cqn14 and EDASeq15 could not remove these false positives, likely because GC content bias is not the only source that contributes to bias in gene expression data.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
In the citing paper
“This confirms the impact of two already recognized RNA-seq latent variables, GC-content and insert size15,22, and it also identifies two more relevant contributions to technical variation, gene coverage variation and error rate.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Controlling the False Discovery Rate: A Practical and Powerful Approach to Multiple Testing
Benjamini, Y., & Hochberg, Y. (1995) · Journal of the Royal Statistical Society Series B: Statistical Methodology · doi:10.1111/j.2517-6161.1995.tb02031.x · Cited by 109,482
Robinson, M. D., McCarthy, D. J., & Smyth, G. K. (2010) · Bioinformatics · doi:10.1093/bioinformatics/btp616 · Cited by 41,364
Ultrafast and memory-efficient alignment of short DNA sequences to the human genome.
Langmead, B., Trapnell, C., Pop, M., & Salzberg, S. L. (2009) · Genome Biology · doi:10.1186/gb-2009-10-3-r25 · Cited by 21,849
Differential expression analysis for sequence count data
Anders, S., & Huber, W. (2010) · Genome Biology · doi:10.1186/gb-2010-11-10-r106 · Cited by 15,581
Mapping and quantifying mammalian transcriptomes by RNA-Seq
Mortazavi, A., Williams, B. A., McCue, K., Schaeffer, L., & Wold, B. (2008) · Nature Methods · doi:10.1038/nmeth.1226 · Cited by 13,696
RNA-Seq: a revolutionary tool for transcriptomics
Wang, Z., Gerstein, M., & Snyder, M. (2009) · Nature Reviews Genetics · doi:10.1038/nrg2484 · Cited by 13,174
Smyth, G. K. (2004) · Statistical Applications in Genetics and Molecular Biology · doi:10.2202/1544-6115.1027 · Cited by 11,829
Exploration, normalization, and summaries of high density oligonucleotide array probe level data
Irizarry, R. A. (2003) · Biostatistics · doi:10.1093/biostatistics/4.2.249 · Cited by 10,603
A scaling normalization method for differential expression analysis of RNA-seq data
Robinson, M. D., & Oshlack, A. (2010) · Genome Biology · doi:10.1186/gb-2010-11-3-r25 · Cited by 7,876
Gene ontology analysis for RNA-seq: accounting for selection bias
Young, M. D., Wakefield, M. J., Smyth, G. K., & Oshlack, A. (2010) · Genome Biology · doi:10.1186/gb-2010-11-2-r14 · Cited by 7,107
Cited by
Sorted by citations
Moderated estimation of fold change and dispersion for RNA-seq data with DESeq2
Love, M. I., Huber, W., & Anders, S. (2014) · Genome Biology · doi:10.1186/s13059-014-0550-8 · Cited by 95,106
TCGAbiolinks: an R/Bioconductor package for integrative analysis of TCGA data
Colaprico, A., Silva, T. C., Olsen, C., Garofano, L., Cava, C., Garolini, D., Sabedot, T. S., Malta, T. M., Pagnotta, S. M., Castiglioni, I., et al. (2015) · Nucleic Acids Research · doi:10.1093/nar/gkv1507 · Cited by 4,205
A survey of best practices for RNA-seq data analysis
Conesa, A., Madrigal, P., Tarazona, S., Gomez-Cabrero, D., Cervera, A., McPherson, A., Szcześniak, M. W., Gaffney, D. J., Elo, L. L., Zhang, X., et al. (2016) · Genome Biology · doi:10.1186/s13059-016-0881-8 · Cited by 2,794
Normalization of RNA-seq data using factor analysis of control genes or samples
Risso, D., Ngai, J., Speed, T. P., & Dudoit, S. (2014) · Nature Biotechnology · doi:10.1038/nbt.2931 · Cited by 2,052
Self-citationRNA sequencing: the teenage years
Stark, R., Grzelak, M., & Hadfield, J. (2019) · Nature Reviews Genetics · doi:10.1038/s41576-019-0150-2 · Cited by 2,041
Dillies, M., Rau, A., Aubert, J., Hennequet‐Antier, C., Jeanmougin, M., Servant, N., Keime, C., Marot, G., Castel, D., Estellé, J., et al. (2012) · Briefings in Bioinformatics · doi:10.1093/bib/bbs046 · Cited by 1,363
Count-based differential expression analysis of RNA sequencing data using R and Bioconductor
Anders, S., McCarthy, D. J., Chen, Y., Okoniewski, M. J., Smyth, G. K., Huber, W. K., & Robinson, M. D. (2013) · Nature Protocols · doi:10.1038/nprot.2013.099 · Cited by 1,170
Monaco, G., Lee, B., Xu, W., Mustafah, S., Hwang, Y. Y., Carré, C., Burdin, N., Visan, L., Ceccarelli, M., Poidinger, M., et al. (2019) · Cell Reports · doi:10.1016/j.celrep.2019.01.041 · Cited by 958
Comparison of RNA-Seq and Microarray in Transcriptome Profiling of Activated T Cells
Zhao, S., Fung‐Leung, W., Bittner, A., Ngo, K., & Liu, X. (2014) · PLoS ONE · doi:10.1371/journal.pone.0078644 · Cited by 1,000
Chen, Y., Chen, L., Lun, A. T. L., Baldoni, P. L., & Smyth, G. K. (2025) · Nucleic Acids Research · doi:10.1093/nar/gkaf018 · Cited by 951
RobiNA: a user-friendly, integrated software solution for RNA-Seq-based transcriptomics
Lohse, M., Bolger, A., Nagel, A., Fernie, A. R., Lunn, J. E., Stitt, M., & Usadel, B. (2012) · Nucleic Acids Research · doi:10.1093/nar/gks540 · Cited by 790
Data quality aware analysis of differential expression in RNA-seq with NOISeq R/Bioc package
Tarazona, S., Furió‐Tarí, P., Turrà, D., Pietro, A. D., Nueda, M. J., Ferrer, A., & Conesa, A. (2015) · Nucleic Acids Research · doi:10.1093/nar/gkv711 · Cited by 787
variancePartition: interpreting drivers of variation in complex gene expression studies
Hoffman, G. E., & Schadt, E. E. (2016) · BMC Bioinformatics · doi:10.1186/s12859-016-1323-z · Cited by 769
A general and flexible method for signal extraction from single-cell RNA-seq data
Risso, D., Perraudeau, F., Gribkova, S., Dudoit, S., & Vert, J.-P. (2018) · Nature Communications · doi:10.1038/s41467-017-02554-5 · Cited by 718
Self-citationRevealing the vectors of cellular identity with single-cell genomics
Wagner, A., Regev, A., & Yosef, N. (2016) · Nature Biotechnology · doi:10.1038/nbt.3711 · Cited by 632
Comprehensive analysis of normal adjacent to tumor transcriptomes
Aran, D., Camarda, R., Odegaard, J. I., Paik, H., Oskotsky, B., Krings, G., Goga, A., Sirota, M., & Butte, A. J. (2017) · Nature Communications · doi:10.1038/s41467-017-01027-z · Cited by 620
Computational analysis of bacterial RNA-Seq data
McClure, R., Balasubramanian, D., Sun, Y., Bobrovskyy, M., Sumby, P., Genco, C. A., Vanderpool, C. K., & Tjaden, B. C. (2013) · Nucleic Acids Research · doi:10.1093/nar/gkt444 · Cited by 584
Systems vaccinology of the BNT162b2 mRNA vaccine in humans
Arunachalam, P. S., Scott, M. K. D., Hagan, T., Li, C., Feng, Y., Wimmers, F., Grigoryan, L., Trisal, M., Edara, V. V., Lai, L., et al. (2021) · Nature · doi:10.1038/s41586-021-03791-x · Cited by 572
Mounir, M., Lucchetta, M., Silva, T. C., Olsen, C., Bontempi, G., Chen, X., Noushmehr, H., Colaprico, A., & Papaleo, E. (2019) · PLoS Computational Biology · doi:10.1371/journal.pcbi.1006701 · Cited by 535
Gupta, S., Ellis, S. E., Ashar, F. N., Moes, A., Bader, J. S., Zhan, J., West, A. B., & Arking, D. E. (2014) · Nature Communications · doi:10.1038/ncomms6748 · Cited by 504
“…the gene count data was normalized for within and between lane biases (e.g. GC content) and sequencing depth by methods implemented in Conditional Quantile Normalization 45 (CQN) and Exploratory Data Analysis and Normalization for RNA-Seq (EDASeq) 46 , using the default settings for each method.”
A new view of transcriptome complexity and regulation through the lens of local splicing variations
Vaquero-Garcia, J., Barrera, A., Gazzara, M. R., González-Vallinas, J., Lahens, N. F., Hogenesch, J. B., Lynch, K. W., & Barash, Y. (2016) · eLife · doi:10.7554/elife.11752 · Cited by 488
Design and computational analysis of single-cell RNA-sequencing experiments
Bacher, R. L., & Kendziorski, C. M. (2016) · Genome biology · doi:10.1186/s13059-016-0927-y · Cited by 460
Generation of a Broadly Useful Model for COVID-19 Pathogenesis, Vaccination, and Treatment
Sun, J., Zhuang, Z., Zheng, J. H., Li, K., Wong, R. L.-Y., Liu, D., Huang, J., He, J., Zhu, A., Zhao, J., et al. (2020) · Cell · doi:10.1016/j.cell.2020.06.010 · Cited by 440
Necroptosis microenvironment directs lineage commitment in liver cancer
Seehawer, M., Heinzmann, F., D’Artista, L., Harbig, J., Roux, P.-F., Hoenicke, L., Dang, H., Klotz, S., Robinson, L., Doré, G., et al. (2018) · Nature · doi:10.1038/s41586-018-0519-y · Cited by 406
RNA-Seq workflow: gene-level exploratory analysis and differential expression
Love, M. I., Anders, S., Kim, V., & Huber, W. K. (2015) · F1000Research · doi:10.12688/f1000research.7035.1 · Cited by 404
Genetic improvement of tomato by targeted control of fruit softening
Uluışık, S., Chapman, N. H., Smith, R. A., Poole, M., Adams, G. G., Gillis, R. B., Besong, T. M. D., Sheldon, J. M., Stiegelmeyer, S. M., Pérez-Fons, L., et al. (2016) · Nature Biotechnology · doi:10.1038/nbt.3602 · Cited by 391
Selecting between-sample RNA-Seq normalization methods from the perspective of their assumptions
Evans, C., Hardin, J., & Stoebel, D. M. (2017) · Briefings in Bioinformatics · doi:10.1093/bib/bbx008 · Cited by 368
Garofano, L., Migliozzi, S., Oh, Y. T., D’Angelo, F., Najac, R. D., Ko, A., Frangaj, B., Caruso, F. P., Yu, K., Yuan, J., et al. (2021) · Nature Cancer · doi:10.1038/s43018-020-00159-4 · Cited by 380
Transcriptomics in the RNA-seq era
McGettigan, P. (2013) · Current Opinion in Chemical Biology · doi:10.1016/j.cbpa.2012.12.008 · Cited by 383
Next maSigPro: updating maSigPro bioconductor package for RNA-seq time series
Nueda, M. J., Tarazona, S., & Conesa, A. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu333 · Cited by 348
Atlas of clinically distinct cell states and ecosystems across human solid tumors
Luca, B. A., Steen, C. B., Matusiak, M., Azizi, A., Varma, S., Zhu, C., Przybył, J., Espín-Pérez, A., Diehn, M., Alizadeh, A. A., et al. (2021) · Cell · doi:10.1016/j.cell.2021.09.014 · Cited by 359
Swarm Intelligence-Enhanced Detection of Non-Small-Cell Lung Cancer Using Tumor-Educated Platelets
Best, M. G., Sol, N., Veld, S. G. J. G. I. ‘., Vancura, A., Muller, M., Niemeijer, A.-L. N., Fejes, A. V., Tjon‐Kon‐Fat, L., Veld, A. E. H. I. ‘., Leurs, C. E., et al. (2017) · Cancer Cell · doi:10.1016/j.ccell.2017.07.004 · Cited by 341
Multifunctional role of the transcription factor Blimp-1 in coordinating plasma cell differentiation
Minnich, M., Tagoh, H., Bönelt, P., Axelsson, E., Fischer, M., Cebolla, B., Tarakhovsky, A., Nutt, S. L., Jaritz, M., & Busslinger, M. (2016) · Nature Immunology · doi:10.1038/ni.3349 · Cited by 335
Polyester : simulating RNA-seq datasets with differential transcript expression
Frazee, A. C., Jaffe, A. E., Langmead, B., & Leek, J. T. (2015) · Bioinformatics · doi:10.1093/bioinformatics/btv272 · Cited by 313
TCGA Workflow: Analyze cancer genomics and epigenomics data using Bioconductor packages
Silva, T. C., Colaprico, A., Olsen, C., D’Angelo, F., Bontempi, G., Ceccarelli, M., & Noushmehr, H. (2016) · F1000Research · doi:10.12688/f1000research.8923.1 · Cited by 303
Franzén, O., Ermel, R., Cohain, A., Akers, N. K., Di Narzo, A., Talukdar, H. A., Foroughi-Asl, H., Giambartolomei, C., Fullard, J. F., Sukhavasi, K., et al. (2016) · Science · doi:10.1126/science.aad6970 · Cited by 288
On the length, weight and GC content of the human genome
Piovesan, A., Pelleri, M. C., Antonaros, F., Strippoli, P., Caracausi, M., & Vitale, L. (2019) · BMC Research Notes · doi:10.1186/s13104-019-4137-z · Cited by 283
Lehmann, B. D., Colaprico, A., Silva, T. C., Chen, J., An, H., Ban, Y. J., Huang, H., Wang, L. H.-C., James, J. L., Balko, J. M., et al. (2021) · Nature Communications · doi:10.1038/s41467-021-26502-6 · Cited by 298
SCnorm: robust normalization of single-cell RNA-seq data
Bacher, R., Chu, L.-F., Leng, N., Gasch, A. P., Thomson, J. A., Stewart, R. M., Newton, M., & Kendziorski, C. (2017) · Nature Methods · doi:10.1038/nmeth.4263 · Cited by 282
Advanced Applications of RNA Sequencing and Challenges
Han, Y., Gao, S., Muegge, K., Zhang, W., & Zhou, B. (2015) · Bioinformatics and Biology Insights · doi:10.4137/bbi.s28991 · Cited by 303
A systematic evaluation of single cell RNA-seq analysis pipelines
Vieth, B., Parekh, S., Ziegenhain, C., Enard, W., & Hellmann, I. (2019) · Nature Communications · doi:10.1038/s41467-019-12266-7 · Cited by 265
Data-based filtering for replicated high-throughput transcriptome sequencing experiments
Rau, A., Gallopin, M., Celeux, G., & Jaffrézic, F. (2013) · Bioinformatics · doi:10.1093/bioinformatics/btt350 · Cited by 262
Human iPSC Glial Mouse Chimeras Reveal Glial Contributions to Schizophrenia
Windrem, M. S., Osipovitch, M., Liu, Z., Bates, J., Chandler-Militello, D., Zou, L., Munir, J., Schanz, S. J., McCoy, K., Miller, R. H., et al. (2017) · Cell stem cell · doi:10.1016/j.stem.2017.06.012 · Cited by 259
Chromatin Accessibility Dynamics during iPSC Reprogramming
Li, D., Liu, J., Yang, X., Zhou, C., Guo, J., Wu, C., Qin, Y., Guo, L., He, J., Yu, S., et al. (2017) · Cell Stem Cell · doi:10.1016/j.stem.2017.10.012 · Cited by 251
Reproducibility of high-throughput mRNA and small RNA sequencing across laboratories
The GEUVADIS Consortium, 't Hoen, P. A. C., Friedländer, M. R., Almlöf, J., Sammeth, M., Pulyakhina, I., Anvar, S. Y., Laros, J. F. J., Buermans, H. P. J., Karlberg, O., et al. (2013) · Nature Biotechnology · doi:10.1038/nbt.2702 · Cited by 250
A new shrinkage estimator for dispersion improves differential expression detection in RNA-seq data
Wu, H., Wang, C., & Wu, Z. J. (2012) · Biostatistics · doi:10.1093/biostatistics/kxs033 · Cited by 248
Li, S., Tighe, S. W., Nicolet, C. M., Grove, D., Levy, S., Farmerie, W., Viale, A., Wright, C., Schweitzer, P. A., Gao, Y., et al. (2014) · Nature Biotechnology · doi:10.1038/nbt.2972 · Cited by 240
MNase titration reveals differences between nucleosome occupancy and chromatin accessibility
Mieczkowski, J., Cook, A., Bowman, S., Mueller, B., Alver, B. H., Kundu, S., Deaton, A. M., Urban, J. A., Larschan, E., Park, P. J., et al. (2016) · Nature Communications · doi:10.1038/ncomms11485 · Cited by 235
Measuring differential gene expression with RNA-seq: challenges and strategies for data analysis
Finotello, F., & Camillo, B. D. (2014) · Briefings in Functional Genomics · doi:10.1093/bfgp/elu035 · Cited by 253
Nicolás, C., Martin-Bertelsen, T., Floudas, D., Bentzer, J., Smits, M. M., Johansson, T., Troein, C., Persson, P., & Tunlid, A. (2018) · The ISME Journal · doi:10.1038/s41396-018-0331-6 · Cited by 229
Trimming of sequence reads alters RNA-Seq gene expression estimates
Williams, C., Baccarella, A., Parrish, J. Z., & Kim, C. C. (2016) · BMC Bioinformatics · doi:10.1186/s12859-016-0956-2 · Cited by 231
Carpinelli, E. C., Telatin, A., Vitulo, N., Forcato, C., D’Angelo, M., Schiavon, R., Vezzi, A., Giacometti, G. M., Morosinotto, T., & Valle, G. (2013) · Molecular Plant · doi:10.1093/mp/sst120 · Cited by 218
The molecular landscape of glioma in patients with Neurofibromatosis 1
D’Angelo, F., Ceccarelli, M., Tala, Garofano, L., Zhang, J., Frattini, V., Caruso, F. P., Lewis, G., Alfaro-Munoz, K. D., Bauchet, L., et al. (2018) · Nature Medicine · doi:10.1038/s41591-018-0263-8 · Cited by 220
Robles, J. A., Qureshi, S. E., Stephen, S. J., Wilson, S. R., Burden, C. J., & Taylor, J. M. (2012) · BMC Genomics · doi:10.1186/1471-2164-13-484 · Cited by 198
He, J., Babarinde, I. A., Sun, L., Xu, S., Chen, R., Shi, J., Wei, Y., Li, Y., Ma, G., Zhuang, Q., et al. (2021) · Nature Communications · doi:10.1038/s41467-021-21808-x · Cited by 205
Romero, I. G., Pavlovic, B. J., Hernando-Herraez, I., Zhou, X., Ward, M. C., Banovich, N. E., Kagan, C. L., Burnett, J. E., Huang, C. H., Mitrano, A., et al. (2015) · eLife · doi:10.7554/elife.07103 · Cited by 197
A metabolic function of FGFR3-TACC3 gene fusions in cancer
Frattini, V., Pagnotta, S. M., Tala, Fan, J. J., Russo, M. V., Lee, S. B., Garofano, L., Zhang, J., Shi, P., Lewis, G., et al. (2018) · Nature · doi:10.1038/nature25171 · Cited by 189
Love, M. I., Hogenesch, J. B., & Irizarry, R. A. (2016) · Nature Biotechnology · doi:10.1038/nbt.3682 · Cited by 186
HTR7 Mediates Serotonergic Acute and Chronic Itch
Morita, T., McClain, S. P., Batia, L. M., Pellegrino, M., Wilson, S. R., Kienzler, M. A., Lyman, K. A., Olsen, A. S. B., Wong, J. F., Stucky, C. L., et al. (2015) · Neuron · doi:10.1016/j.neuron.2015.05.044 · Cited by 188
Genomic landscape of carcinogen-induced and genetically induced mouse skin squamous cell carcinoma
Nassar, D., Latil, M., Boeckx, B., Lambrechts, D., & Blanpain, C. (2015) · Nature Medicine · doi:10.1038/nm.3878 · Cited by 188
Detecting and correcting systematic variation in large-scale RNA sequencing data
Li, S., Łabaj, P. P., Zumbo, P., Sykacek, P., Shi, W., Shi, L., Phan, J., Wu, P.-Y., Wang, M., Wang, C., et al. (2014) · Nature Biotechnology · doi:10.1038/nbt.3000 · Cited by 175
Comprehensive multi-center assessment of small RNA-seq methods for quantitative miRNA profiling
Giraldez, M. D., Spengler, R. M., Etheridge, A., Godoy, P. M., Barczak, A. J., Srinivasan, S., De Hoff, P. L., Tanriverdi, K., Courtright, A., Lu, S., et al. (2018) · Nature Biotechnology · doi:10.1038/nbt.4183 · Cited by 177
Performance Assessment and Selection of Normalization Procedures for Single-Cell RNA-Seq
Cole, M. B., Risso, D., Wagner, A., DeTomaso, D., Ngai, J., Purdom, E., Dudoit, S., & Yosef, N. (2019) · Cell Systems · doi:10.1016/j.cels.2019.03.010 · Cited by 173
Self-citationCieslik, M. P., Chugh, R. K., Wu, Y.-M., Wu, M., Brennan, C. A., Lonigro, R. J., Su, F., Wang, R., Siddiqui, J., Mehra, R., et al. (2015) · Genome Research · doi:10.1101/gr.189621.115 · Cited by 174
Cobolt: integrative analysis of multimodal single-cell sequencing data
Gong, B., Zhou, Y., & Purdom, E. (2021) · Genome Biology · doi:10.1186/s13059-021-02556-z · Cited by 168
Trehalose 6-Phosphate Regulates Photosynthesis and Assimilate Partitioning in Reproductive Tissue
Oszvald, M., Primavesi, L. F., Griffiths, C. A., Cohn, J., Basu, S. S., Nuccio, M. L., & Paul, M. J. (2018) · PLANT PHYSIOLOGY · doi:10.1104/pp.17.01673 · Cited by 170
De Smedt, L., Palmans, S., Andel, D., Govaere, O., Boeckx, B., Smeets, D., Galle, E., Wouters, J., Barras, D., Suffiotti, M., et al. (2017) · British Journal of Cancer · doi:10.1038/bjc.2016.382 · Cited by 163
RNA-seq and microarray complement each other in transcriptome profiling
Kogenaru, S., Yan, Q., Guo, Y., & Wang, N. (2012) · BMC Genomics · doi:10.1186/1471-2164-13-629 · Cited by 157
Global analysis of mutations driving microevolution of a heterozygous diploid fungal pathogen
Ene, I. V., Farrer, R. A., Hirakawa, M. P., Agwamba, K., Cuomo, C. A., & Bennett, R. J. (2018) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.1806002115 · Cited by 155
Developmental chromatin programs determine oncogenic competence in melanoma
Baggiolini, A., Callahan, S. J., Montal, E. D., Weiss, J. M., Trieu, T., Tagore, M. M., Tischfield, S. E., Walsh, R. M., Suresh, S., Fan, Y., et al. (2021) · Science · doi:10.1126/science.abc1048 · Cited by 154
Gene expression based inference of cancer drug sensitivity
Chawla, S., Rockstroh, A., Lehman, M., Ratther, E., Jain, A., Anand, A., Gupta, A., Bhattacharya, N., Poonia, S., Rai, P., et al. (2022) · Nature Communications · doi:10.1038/s41467-022-33291-z · Cited by 150
RNA-Seq and molecular docking reveal multi-level pesticide resistance in the bed bug
Mamidala, P., Wijeratne, A., Wijeratne, S., Kornacker, K., Sudhamalla, B., Rivera-Vega, L. J., Hoelmer, A., Meulia, T., Jones, S. C., & Mittapalli, O. (2012) · BMC Genomics · doi:10.1186/1471-2164-13-6 · Cited by 148
Enhancer Architecture and Essential Core Regulatory Circuitry of Chronic Lymphocytic Leukemia
Ott, C. J., Alexander Joel Federation, Schwartz, L. S., Kasar, S. N., Klitgaard, J. L., Lenci, R., Li, Q., Lawlor, M. A., Fernandes, S. M., Souza, A., et al. (2018) · Cancer Cell · doi:10.1016/j.ccell.2018.11.001 · Cited by 152
RNA fate determination through cotranscriptional adenosine methylation and microprocessor binding
Knuckles, P., Carl, S. H., Musheev, M. U., Niehrs, C., Wenger, A., & Bühler, M. (2017) · Nature Structural & Molecular Biology · doi:10.1038/nsmb.3419 · Cited by 152
RNA-seq data science: From raw data to effective interpretation
Deshpande, D., Chhugani, K., Chang, Y., Karlsberg, A., Loeffler, C., Zhang, J., Muszyńska, A., Munteanu, V., Yang, H. T., Rotman, J., et al. (2023) · Frontiers in Genetics · doi:10.3389/fgene.2023.997383 · Cited by 158
Atak, Z. K., Gianfelici, V., Hulselmans, G. J., Keersmaecker, K. D., Devasia, A. G., Geerdens, E., Mentens, N., Chiaretti, S., Durinck, K., Uyttebroeck, A., et al. (2013) · PLoS Genetics · doi:10.1371/journal.pgen.1003997 · Cited by 149
Bunnik, E. M., Chung, D.-W. D., Hamilton, M., Ponts, N., Saraf, A., Prudhomme, J. G., Florens, L. A., & Roch, K. G. L. (2013) · Genome biology · doi:10.1186/gb-2013-14-11-r128 · Cited by 146
Kühl, I., Miranda, M., Posse, V., Milenkovic, D., Mourier, A., Siira, S. J., Bonekamp, N. A., Neumann, U., Filipovska, A., Polosa, P. L., et al. (2016) · Science Advances · doi:10.1126/sciadv.1600963 · Cited by 151
RNA Sequencing Data: Hitchhiker's Guide to Expression Analysis
den Berge, K. V., Hembach, K., Soneson, C., Tiberi, S., Clement, L., Love, M. I., Patro, R., & Robinson, M. D. (2019) · Annual Review of Biomedical Data Science · doi:10.1146/annurev-biodatasci-072018-021255 · Cited by 153
Macrophages are metabolically heterogeneous within the tumor microenvironment
Geeraerts, X., Fernández-García, J., Hartmann, F. J., de Goede, K. E., Martens, L., Elkrim, Y., Debraekeleer, A., Stijlemans, B., Vandekeere, A., Rinaldi, G., et al. (2021) · Cell Reports · doi:10.1016/j.celrep.2021.110171 · Cited by 151
Detection and localization of early- and late-stage cancers using platelet RNA
Veld, S. G. J. G. I. ‘., Arkani, M., Post, E., Antunes‐Ferreira, M., D’Ambrosi, S., Vessies, D. C. L., Vermunt, L., Vancura, A., Muller, M., Niemeijer, A.-L. N., et al. (2022) · Cancer Cell · doi:10.1016/j.ccell.2022.08.006 · Cited by 142
He, J., Fu, X., Zhang, M., He, F., Li, W., Abdul, M. M., Zhou, J., Sun, L., Chang, C., Li, Y., et al. (2018) · Nature Communications · doi:10.1038/s41467-018-08006-y · Cited by 145
Humphrey, J., Venkatesh, S., Hasan, R., Herb, J. T., de Paiva Lopes, K., Küçükali, F., Byrska-Bishop, M., Evani, U. S., Narzisi, G., Fagegaltier, D., et al. (2023) · Nature Neuroscience · doi:10.1038/s41593-022-01205-3 · Cited by 138
Acidophilic green algal genome provides insights into adaptation to an acidic environment
Hirooka, S., Hirose, Y., Kanesaki, Y., Higuchi, S., Fujiwara, T., Onuma, R., Era, A., Ohbayashi, R., Uzuka, A., Nozaki, H., et al. (2017) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.1707072114 · Cited by 144
Identification of evolutionarily conserved gene networks mediating neurodegenerative dementia
Swarup, V., Hinz, F. I., Rexach, J. E., Noguchi, K.-I., Toyoshiba, H., Oda, A., Hirai, K., Sarkar, A., Seyfried, N. T., Cheng, C., et al. (2019) · Nature Medicine · doi:10.1038/s41591-018-0223-3 · Cited by 136
Comparison of normalization methods for differential gene expression analysis in RNA-Seq experiments
Maza, É., Frasse, P., Senin, P., Bouzayen, M., & Zouine, M. (2013) · Communicative & Integrative Biology · doi:10.4161/cib.25849 · Cited by 145
Alboresi, A., Perin, G., Vitulo, N., Diretto, G., Block, M. A., Jouhet, J., Meneghesso, A., Valle, G., Giuliano, G., Maréchal, É., et al. (2016) · PLANT PHYSIOLOGY · doi:10.1104/pp.16.00599 · Cited by 139
Li, Q., Hutchins, A. P., Chen, Y., Li, S., Shan, Y., Liao, B., Zheng, D., Shi, X., Li, Y., Chan, W., et al. (2017) · Nature Communications · doi:10.1038/ncomms15166 · Cited by 129
Impulse model-based differential expression analysis of time course sequencing data
Fischer, D. S., Theis, F. J., & Yosef, N. (2018) · Nucleic Acids Research · doi:10.1093/nar/gky675 · Cited by 126
Aging is associated with a systemic length-associated transcriptome imbalance
Stoeger, T., Grant, R. A., McQuattie‐Pimentel, A. C., Anekalla, K. R., Liu, S. S., Navarro, H. T., Singer, B. D., Abdala‐Valencia, H., Schwake, M., Tétreault, M., et al. (2022) · Nature Aging · doi:10.1038/s43587-022-00317-6 · Cited by 129
Gene-by-environment interactions in urban populations modulate risk phenotypes
Favé, M.-J., Lamaze, F. C., Soave, D. M., Hodgkinson, A., Gauvin, H., Bruat, V., Grenier, J., Gbeha, E., Skead, K., Smargiassi, A., et al. (2018) · Nature Communications · doi:10.1038/s41467-018-03202-2 · Cited by 125
A new normalization for Nanostring nCounter gene expression data
Molania, R., Gagnon-Bartsch, J. A., Dobrovic, A., & Speed, T. P. (2019) · Nucleic Acids Research · doi:10.1093/nar/gkz433 · Cited by 123
Probabilistic error correction for RNA sequencing
Le, H.-S., Schulz, M. H., McCauley, B. S., Hinman, V. F., & Bar‐Joseph, Z. (2013) · Nucleic Acids Research · doi:10.1093/nar/gkt215 · Cited by 125
Multi-perspective quality control of Illumina RNA sequencing data analysis
Sheng, Q., Vickers, K. C., Zhao, S., Wang, J., Samuels, D. C., Koues, O. I., Shyr, Y., & Guo, Y. P. (2016) · Briefings in Functional Genomics · doi:10.1093/bfgp/elw035 · Cited by 123
Predictive models of subcellular localization of long RNAs
Zuckerman, B. M., & Ulitsky, I. (2019) · RNA · doi:10.1261/rna.068288.118 · Cited by 124
RNA-Seq Perspectives to Improve Clinical Diagnosis
Marco-Puche, G., Lois, S., Benitez, J. J., & Triviño, J. C. (2019) · Frontiers in Genetics · doi:10.3389/fgene.2019.01152 · Cited by 119
A mouse-specific retrotransposon drives a conserved Cdk2ap1 isoform essential for development
Modzelewski, A. J., Shao, W., Chen, J., Lee, A., Qi, X., Noon, M., Tjokro, K., Sales, G., Biton, A., Anand, A., et al. (2021) · Cell · doi:10.1016/j.cell.2021.09.021 · Cited by 120
Self-citationCoNekT: an open-source framework for comparative genomic and transcriptomic network analyses
Proost, S., & Mutwil, M. (2018) · Nucleic Acids Research · doi:10.1093/nar/gky336 · Cited by 113
Wilson, J. A. C., Prow, N. A., Schroder, W. A., Ellis, J. J., Cumming, H. E., Gearing, L. J., Poo, Y. S., Taylor, A., Hertzog, P. J., Giallonardo, F. D., et al. (2017) · PLoS Pathogens · doi:10.1371/journal.ppat.1006155 · Cited by 116
Engelholm, L. H., Riaz, A., Serra, D., Dagnæs‐Hansen, F., Johansen, J. V., Santoni‐Rugiu, E., Hansen, S. H., Niola, F., & Frödin, M. (2017) · Gastroenterology · doi:10.1053/j.gastro.2017.09.008 · Cited by 112
Metadata
- Authors
- Risso, D., Schwartz, K., Sherlock, G., & Dudoit, S.
- Affiliations
- University of Padua (IT)
- Stanford University (US)
- University of California, Berkeley (US)
- Corresponding author
- Dudoit, Sandrine
- Venue
- BMC Bioinformatics
- Published
- 2011-12
- Volume
- 12
- Issue
- 1
- Pages
- 480-480
- ISSN
- 1471-2105 (ISSN-L)
- PMC
- PMC3315510
- OpenAlex
- W2168342464
- Cited by
- 867
- References
- 35
- Citations by year
- 2026: 29 · 2025: 58 · 2024: 83 · 2023: 80 · 2022: 89 · 2021: 97 · 2020: 74 · 2019: 55 · 2018: 60 · 2017: 49 · 2016: 59 · 2015: 38 · 2014: 48 · 2013: 30 · 2012: 18
- FWCI
- 12.48
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- cc-by
- Locations
- BMC Bioinformatics (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Collection of Biostatistics Research Archive (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- eScholarship (California Digital Library) (submittedVersion)
- Padua Research Archive (University of Padova) (submittedVersion, cc-by)
- Funders
- Università degli Studi di Padova
- National Institutes of Health
- National Human Genome Research Institute (R01 HG03468, R01HG003468)
- National Institute of Allergy and Infectious Diseases (R01AI077737)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene expression and cancer classification
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Keywords
- RNA-seq, differential gene expression, gene expression bias, transcriptome sequencing
- MeSH terms
- Base Composition, Saccharomyces cerevisiae, Sequence Analysis, RNA, Gene Expression Profiling, Transcriptome
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
