

The DOI matches its Crossref record; 37 of the 37 references it cites were found in a registry or an index.
GC-Content Normalization for RNA-Seq Data
Open accessAbstract
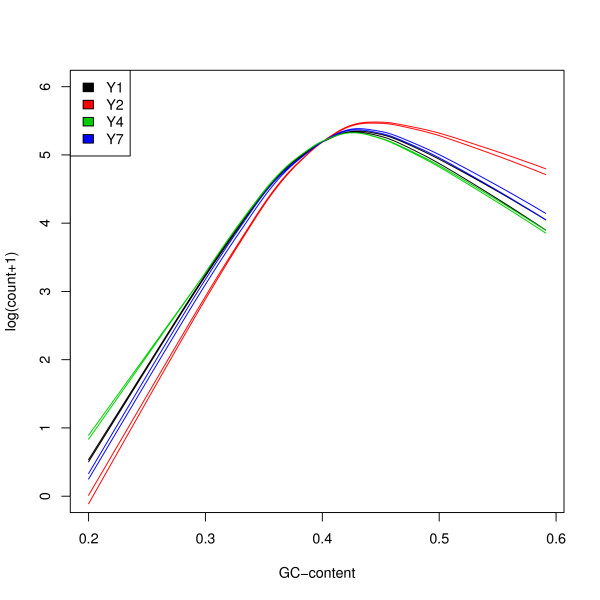
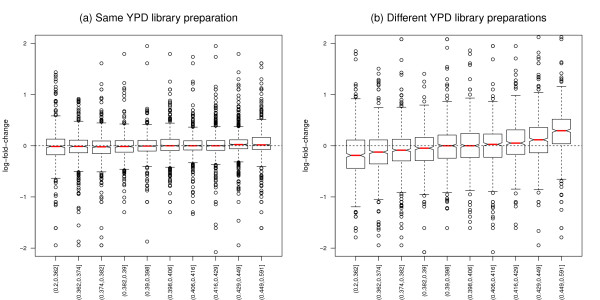
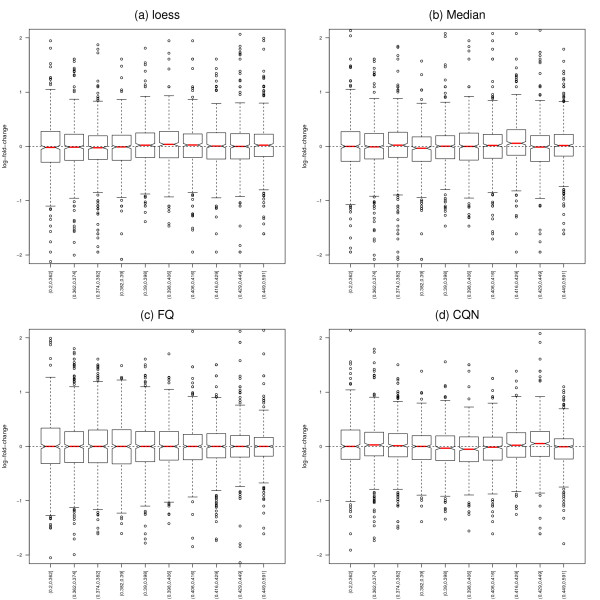
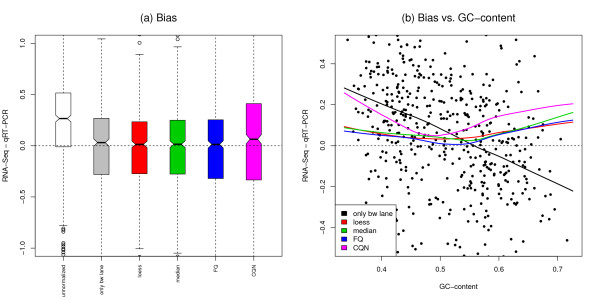
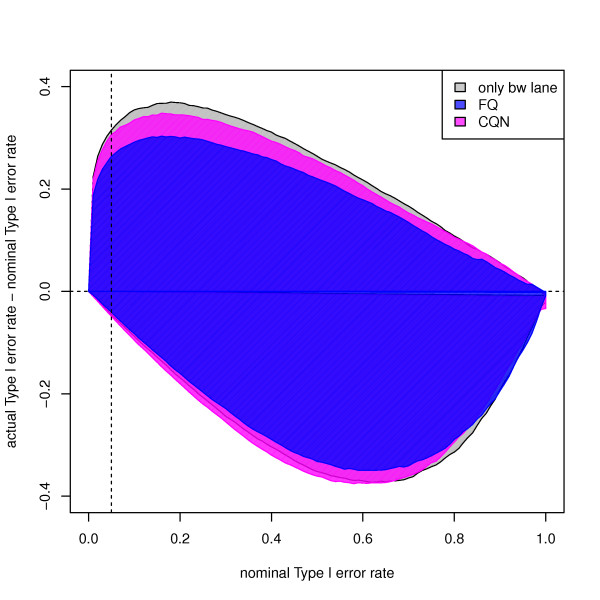
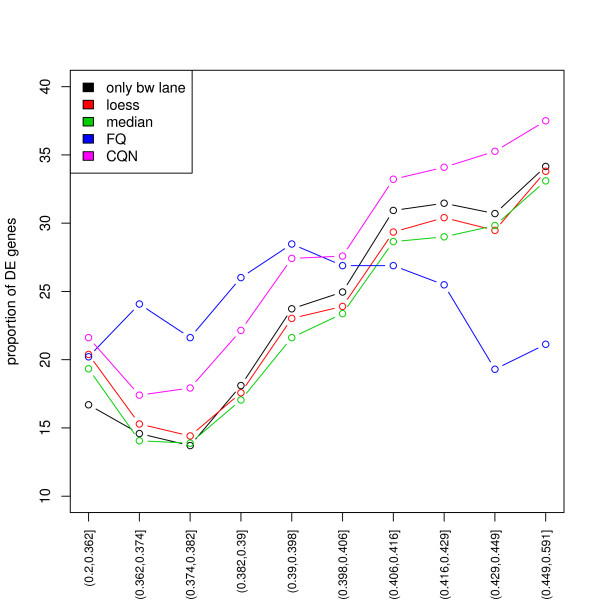
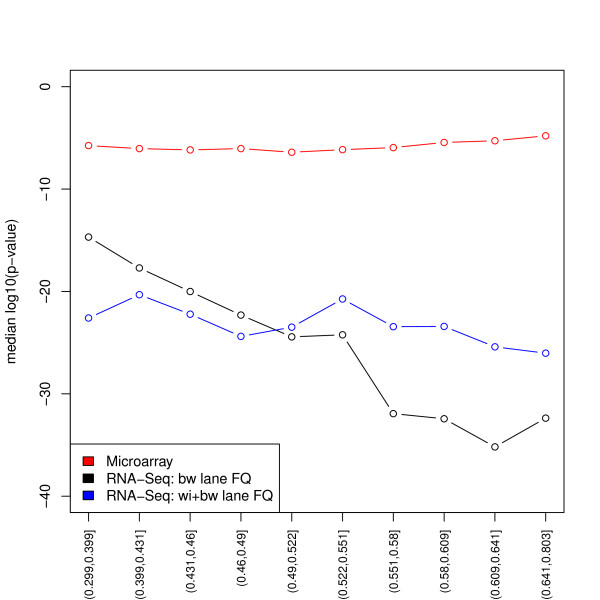
Background: Transcriptome sequencing (RNA-Seq) has become the assay of choice for high-throughput studies of gene expression. However, as is the case with microarrays, major technology-related artifacts and biases affect the resulting expression measures. Normalization is therefore essential to ensure accurate inference of expression levels and subsequent analyses thereof. Results: We focus on biases related to GC-content and demonstrate the existence of strong sample-specific GC-content effects on RNA-Seq read counts, which can substantially bias differential expression analysis. We propose three simple within-lane gene-level GC-content normalization approaches and assess their performance on two different RNA-Seq datasets, involving different species and experimental designs. Our methods are compared to state-of-the-art normalization procedures in terms of bias and mean squared error for expression fold-change estimation and in terms of Type I error and p-value distributions for tests of differential expression. The exploratory data analysis and normalization methods proposed in this article are implemented in the open-source Bioconductor R package EDASeq. Conclusions: Our within-lane normalization procedures, followed by between-lane normalization, reduce GC-content bias and lead to more accurate estimates of expression fold-changes and tests of differential expression. Such results are crucial for the biological interpretation of RNA-Seq experiments, where downstream analyses can be sensitive to the supplied lists of genes.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“commonly used methods fornormalization (cqn, EDASeq, RUV2, sva, PEER) varied in their ability to remove these systematic biases, depending on sample complexity and initial data quality.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
Experimental — accuracy not yet measuredIn the citing paper
“These effects can be minimized by appropriate experimental design [51] or, alternatively, removed by batch-correction methods such as COMBAT [52] or ARSyN [20, 53].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“Filtering out genes that are expressed at low levels prior to differential expression analysis reduces the severity of the correction and may improve the power of detection [20].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“NOISeq and NOISeqBIO (the adaptation of NOISeq for biological replication) are more efficient in avoiding false positive calls at the cost of some sensitivity but perform well with different numbers of replicates [10, 20, 212].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“SAMseq performs well in terms of FDR but presents an acceptable sensitivity when the number of replicates is relatively high, at least 10 [20, 55, 209].”
From A survey of best practices for RNA-seq data analysis (2016) · cites this paper as [20]
In the citing paper
“Notably, neither applying GC bias correction tools (cqn14 and EDASeq15) to individual sites nor changing read counts to only use 3′-UTRs was effective at decreasing the number of inter-site false positive DEGs (Fig. 1b and Supplementary Fig. 3); in most cases, these methods actually increased the number of false positives.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
In the citing paper
“The application of GC content bias correction packages including cqn14 and EDASeq15 could not remove these false positives, likely because GC content bias is not the only source that contributes to bias in gene expression data.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
In the citing paper
“This confirms the impact of two already recognized RNA-seq latent variables, GC-content and insert size15,22, and it also identifies two more relevant contributions to technical variation, gene coverage variation and error rate.”
From Detecting and correcting systematic variation in large-scale RNA sequencing data (2014) · cites this paper as [15]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Controlling the False Discovery Rate: A Practical and Powerful Approach to Multiple Testing
Benjamini, Y., & Hochberg, Y. (1995) · Journal of the Royal Statistical Society Series B: Statistical Methodology · doi:10.1111/j.2517-6161.1995.tb02031.x · Cited by 109,482
Robinson, M. D., McCarthy, D. J., & Smyth, G. K. (2010) · Bioinformatics · doi:10.1093/bioinformatics/btp616 · Cited by 41,364
Ultrafast and memory-efficient alignment of short DNA sequences to the human genome.
Langmead, B., Trapnell, C., Pop, M., & Salzberg, S. L. (2009) · Genome Biology · doi:10.1186/gb-2009-10-3-r25 · Cited by 21,849
Differential expression analysis for sequence count data
Anders, S., & Huber, W. (2010) · Genome Biology · doi:10.1186/gb-2010-11-10-r106 · Cited by 15,581
Mapping and quantifying mammalian transcriptomes by RNA-Seq
Mortazavi, A., Williams, B. A., McCue, K., Schaeffer, L., & Wold, B. (2008) · Nature Methods · doi:10.1038/nmeth.1226 · Cited by 13,696
RNA-Seq: a revolutionary tool for transcriptomics
Wang, Z., Gerstein, M., & Snyder, M. (2009) · Nature Reviews Genetics · doi:10.1038/nrg2484 · Cited by 13,174
Smyth, G. K. (2004) · Statistical Applications in Genetics and Molecular Biology · doi:10.2202/1544-6115.1027 · Cited by 11,829
Exploration, normalization, and summaries of high density oligonucleotide array probe level data
Irizarry, R. A. (2003) · Biostatistics · doi:10.1093/biostatistics/4.2.249 · Cited by 10,603
A scaling normalization method for differential expression analysis of RNA-seq data
Robinson, M. D., & Oshlack, A. (2010) · Genome Biology · doi:10.1186/gb-2010-11-3-r25 · Cited by 7,876
Gene ontology analysis for RNA-seq: accounting for selection bias
Young, M. D., Wakefield, M. J., Smyth, G. K., & Oshlack, A. (2010) · Genome Biology · doi:10.1186/gb-2010-11-2-r14 · Cited by 7,107
Cited by
Sorted by citations
Items 801–865 of 865
Additional file 6 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424928
Costa, M. C., Angelini, C., Franzese, M., Iside, C., Salvatore, M., Laezza, L., Napolitano, F., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26693928
Costa, M. C., Angelini, C., Franzese, M., Iside, C., Salvatore, M., Laezza, L., Napolitano, F., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26693913
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716609
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716618
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716591
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716597
Caruso, F. P., D'Andrea, M. R., Coppola, L., Landriscina, M., Condelli, V., Cerulo, L., Giordano, G., Porrás, A., & Pancione, M. (2022) · Figshare · doi:10.6084/m9.figshare.20479638
Additional file 3 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424916
Additional file 4 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424922
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716612
Dittlau, K. S., Terrie, L., Baatsen, P., Kerstens, A., Swert, L. D., Janky, R., Corthout, N., Masrori, P., Damme, P. V., Hyttel, P., et al. (2024) · Figshare · doi:10.6084/m9.figshare.25458004
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716624
Liang, L., He, M., Zhang, Y., Wang, C., Qin, Z., Li, Q., Yang, T., Meng, F., Zhou, Y., Ge, H., et al. (2024) · Figshare · doi:10.6084/m9.figshare.26647368
Costa, M. C., Angelini, C., Franzese, M., Iside, C., Salvatore, M., Laezza, L., Napolitano, F., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26693922
Additional file 1 of Altered hippocampal transcriptome dynamics following sleep deprivation
Gaine, M. E., Bahl, E., Chatterjee, S., Michaelson, J. J., Abel, T. G., & Lyons, L. C. (2021) · Figshare · doi:10.6084/m9.figshare.15161928
Osabe, T., Shimizu, K., & Kadota, K. (2021) · Figshare · doi:10.6084/m9.figshare.16841064
Simeone, I., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26564910
Costa, M. C., Angelini, C., Franzese, M., Iside, C., Salvatore, M., Laezza, L., Napolitano, F., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26693925
Costa, M. C., Angelini, C., Franzese, M., Iside, C., Salvatore, M., Laezza, L., Napolitano, F., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26693916
Moskovicz, V., Ben-El, R. T. G., Horev, G., & Mizrahi, B. (2021) · Figshare · doi:10.6084/m9.figshare.16305939
Osabe, T., Shimizu, K., & Kadota, K. (2021) · Figshare · doi:10.6084/m9.figshare.16841055
Additional file 2 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424913
Caruso, F. P., D'Andrea, M. R., Coppola, L., Landriscina, M., Condelli, V., Cerulo, L., Giordano, G., Porrás, A., & Pancione, M. (2022) · Figshare · doi:10.6084/m9.figshare.20479641
Additional file 1 of Modulation of SLFN11 induces changes in DNA Damage response in breast cancer
Raynaud, C. M., Ahmed, E. I., Jabeen, A., Sanchez, A., Sherif, S., Carneiro-Lobo, T., Awad, A. S., Awartani, D., Naik, A., Thomas, R., et al. (2023) · Figshare · doi:10.6084/m9.figshare.24633304
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.26716615
Liang, L., He, M., Zhang, Y., Wang, C., Qin, Z., Li, Q., Yang, T., Meng, F., Zhou, Y., Ge, H., et al. (2024) · Figshare · doi:10.6084/m9.figshare.24709096
Simeone, I., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26564904
Osabe, T., Shimizu, K., & Kadota, K. (2021) · Figshare · doi:10.6084/m9.figshare.16841052
Simeone, I., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26564913
Liang, L., He, M., Zhang, Y., Wang, C., Qin, Z., Li, Q., Yang, T., Meng, F., Zhou, Y., Ge, H., et al. (2024) · Figshare · doi:10.6084/m9.figshare.26647371
Perycz, M., Dabrowski, M. J., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.25762932
Feng, Y., & Li, L. M. (2021) · Figshare · doi:10.6084/m9.figshare.15071342
Perycz, M., Dąbrowski, M., Jardanowska-Kotuniak, M., Roura, A.-J., Gielniewski, B., Stępniak, K., Dramiński, M., Ciechomska, I. A., Kamińska, B., & Wojtaś, B. (2024) · Figshare · doi:10.6084/m9.figshare.25763097
Additional file 1 of A comparison of transcriptome analysis methods with reference genome
Liu, X., Zhao, J., Xue, L., Zhao, T., Ding, W., Han, Y., & Ye, H. (2022) · Figshare · doi:10.6084/m9.figshare.19424910
Kakati, T., Bhattacharyya, D. K., Kalita, J. K., & Norden‐Krichmar, T. M. (2022) · Figshare · doi:10.6084/m9.figshare.17982047
Simeone, I., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26564919
Osabe, T., Shimizu, K., & Kadota, K. (2021) · Figshare · doi:10.6084/m9.figshare.16841067
Simeone, I., & Ceccarelli, M. (2024) · Figshare · doi:10.6084/m9.figshare.26564916
Lokanc, S., Sipila, P., Cuvelier, G. D. E., Murguía-Favela, L. E., Wright, N. A. M., Grunebaum, E., Bose, P., & Narendran, A. (2026) · Human Immunology · doi:10.1016/j.humimm.2026.111671
Structured factorization for single-cell gene expression data
Canale, A., Galtarossa, L., Risso, D., Schiavon, L., & Toto, G. (2026) · Journal of the Royal Statistical Society Series C (Applied Statistics) · doi:10.1093/jrsssc/qlag009
Self-citationde Freitas, J. A. N. L. F. (2020) · theses.fr (ABES) · doi:10.70675/0e6e3614z3915z4fb8za2cdz24b40b9679b2
Caractérisation de la reprogrammation de l'expression des gènes chez les blés allopyloïdes
Chalabi, S. (2014) · theses.fr (ABES) · doi:10.70675/bee96c14z587dz44d5zad96z53a7d6126cec
Dynamique des éléments transposables et évolution du génome des spartines polyploïdes
Giraud, D. (2019) · theses.fr (ABES) · doi:10.70675/4193c885z9d9az4014z9a91z7219d45c2ab8
MRE11 proximal polyadenylation site-mediated looping impacts transcription and genomic stability
Huang, K., Brault, M. E., Cong, K., Azazmeh, N., Collins, S., Lee, S., Lantz, G. A., Raskind, G., Legube, G., Beroukhim, R., et al. (2026) · Molecular Cell · doi:10.1016/j.molcel.2026.03.032
Lai, W., Wang, L., Tang, C., Chen, D., & Zhong, C. (2026) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.6c00254
Systematic benchmarking of small variant calling pipelines for long-read RNA sequencing dataPreprint
Wang, J., & Robinson, M. D. (2026) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.64898/2026.04.29.721619
Castillo, D. M., & Moyle, L. C. (2026) · Proceedings of the Royal Society B Biological Sciences · doi:10.1098/rspb.2026.0280
Herrgott, G. A., Garofano, L., Morosini, N. S., Done, B., Powell, C., deCarvalho, A. C., Hasselbach, L. A., Transou, A. D., Lee, I. Y., Walbert, T., et al. (2026) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.64898/2026.06.05.730437
Solak, H. C., Mert, N., Leblebici, A., Işık, Z., Ellidokuz, E., & Mert-Ozupek, N. (2026) · PLoS ONE · doi:10.1371/journal.pone.0351228
Wang, K., Huang, X., Gao, H., Li, J., Huang, T., Ke, M., Fu, Y., Lin, Z., Pei, D., & Qin, Y. (2026) · Cell Regeneration · doi:10.1186/s13619-026-00302-z
Shank3 mutation disrupts the molecular signature of sleepiness across developmentPreprint
Wald, E. M., Medina, E., Ottaway, C. A., Muheim, C., Ford, K., Patterson, T., Singletary, K. G., Ingiosi, A. M., & Peixoto, L. (2026) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.64898/2026.08.21.746326
Hu, R., Chen, W., Ding, Z., Zheng, L., (4829), Y. L., Wang, C., Ai, G., Fang, C., He, S., & Yang, L. (2026) · Aquatic Diversity and Ecology · doi:10.1016/j.ade.2026.100024
γδ T cell receptor dependencies define a unique immunosurveillance modality
Veland, N., García-Cassani, B., Zarco-Cuadrillero, Á., Mavrigiannaki, A. M., Natalini, A., Eum, J., Suárez‐Bonnet, A., Vantourout, P., McKenzie, D. R., Franken, J., et al. (2026) · Nature · doi:10.1038/s41586-026-11076-4
Hwangbo, D., Kwon, Y.-J., Iwanaszko, M., Sisobhan, S., Jiang, P., Abbasi, L., Wright, N., Alli, S., Hutchison, A. L., Dinner, A. R., et al. (2026) · eLife · doi:10.7554/elife.86191.2.sa0
Dietary Restriction Impacts Peripheral Diurnal Rhythms Important for Longevity in DrosophilaPreprint
Hwangbo, D., Kwon, Y.-J., Iwanaszko, M., Sisobhan, S., Jiang, P., Abbasi, L., Wright, N., Alli, S., Hutchison, A. L., Dinner, A. R., et al. (2026) · eLife · doi:10.7554/elife.86191.2
Kruawan, A., Stammnitz, M. R., Wilson, R., Ramarathinam, S. H., Ong, C. E. B., Fairfax, K. A., Patchett, A. L., Lyons, A. B., & Flies, A. S. (2026) · Open Biology · doi:10.1098/rsob.250430
Paniagua, A., Jetzinger, F., Tarazona, S., & Conesa, A. (2026) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.64898/2026.10.10.757893
Qi, Y., Miller, L. R., D'Ascenzo, M. D., Berndt, J. D., Whitney, G. A., Duffy, F. J., Danziger, S. A., Peskind, E. R., Li, G., Masters, C. L., et al. (2026) · Scientific Reports · doi:10.1038/s41598-026-74186-z
Metadata
- Authors
- Risso, D., Schwartz, K., Sherlock, G., & Dudoit, S.
- Affiliations
- University of Padua (IT)
- Stanford University (US)
- University of California, Berkeley (US)
- Corresponding author
- Dudoit, Sandrine
- Venue
- BMC Bioinformatics
- Published
- 2011-12
- Volume
- 12
- Issue
- 1
- Pages
- 480-480
- ISSN
- 1471-2105 (ISSN-L)
- PMC
- PMC3315510
- OpenAlex
- W2168342464
- Cited by
- 867
- References
- 35
- Citations by year
- 2026: 29 · 2025: 58 · 2024: 83 · 2023: 80 · 2022: 89 · 2021: 97 · 2020: 74 · 2019: 55 · 2018: 60 · 2017: 49 · 2016: 59 · 2015: 38 · 2014: 48 · 2013: 30 · 2012: 18
- FWCI
- 12.48
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- cc-by
- Locations
- BMC Bioinformatics (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Collection of Biostatistics Research Archive (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- eScholarship (California Digital Library) (submittedVersion)
- Padua Research Archive (University of Padova) (submittedVersion, cc-by)
- Funders
- Università degli Studi di Padova
- National Institutes of Health
- National Human Genome Research Institute (R01 HG03468, R01HG003468)
- National Institute of Allergy and Infectious Diseases (R01AI077737)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene expression and cancer classification
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Keywords
- RNA-seq, differential gene expression, gene expression bias, transcriptome sequencing
- MeSH terms
- Base Composition, Saccharomyces cerevisiae, Sequence Analysis, RNA, Gene Expression Profiling, Transcriptome
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
