

The DOI matches its Crossref record; 111 of the 111 references it cites were found in a registry or an index; 19 of the 40 sources its claims cite were checked for support.
Advancing regulatory variant effect prediction with AlphaGenome
Open accessPublished version of a preprint
Abstract
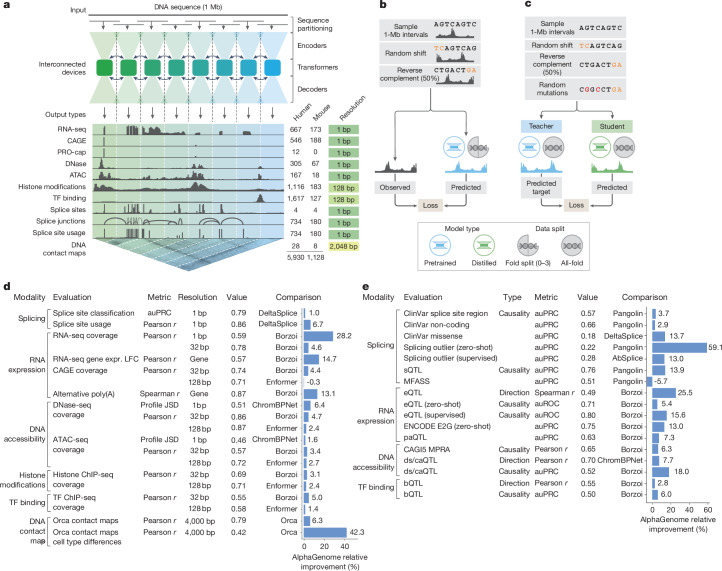
Deep learning models that predict functional genomic measurements from DNA sequences are powerful tools for deciphering the genetic regulatory code. Existing methods involve a trade-off between input sequence length and prediction resolution, thereby limiting their modality scope and performance 1–5 . We present AlphaGenome, a unified DNA sequence model, which takes as input 1 Mb of DNA sequence and predicts thousands of functional genomic tracks up to single-base-pair resolution across diverse modalities. The modalities include gene expression, transcription initiation, chromatin accessibility, histone modifications, transcription factor binding, chromatin contact maps, splice site usage and splice junction coordinates and strength. Trained on human and mouse genomes, AlphaGenome matches or exceeds the strongest available external models in 25 of 26 evaluations of variant effect prediction. The ability of AlphaGenome to simultaneously score variant effects across all modalities accurately recapitulates the mechanisms of clinically relevant variants near the TAL1 oncogene 6 . To facilitate broader use, we provide tools for making genome track and variant effect predictions from sequence.
Overview
AI snapshot
Written from the paper’s full text.
TL;DR
AlphaGenome is a unified DNA sequence-to-function model that takes 1 Mb of DNA and predicts thousands of functional genomic tracks up to single-base-pair resolution across expression, splicing, chromatin, binding and contact-map modalities. It matches or exceeds the strongest external models in 25 of 26 variant effect evaluations and 22 of 24 genome track evaluations, enabling multimodal interpretation of regulatory variants.
Key findings
- AlphaGenome predicts 5,930 human or 1,128 mouse genome tracks across 11 modalities from 1-Mb input, including RNA-seq, CAGE, PRO-cap, splice sites/usage/junctions, DNase, ATAC-seq, histone ChIP-seq, transcription factor binding and contact maps.
- For genome track prediction, AlphaGenome outperformed external models on 22 of 24 evaluations, including +14.7% relative improvement in cell-type-specific gene-level expression log-fold change versus Borzoi, contact map Pearson r +6.3% and cell-type-specific differences +42.3% versus Orca, +15% total counts Pearson r versus ProCapNet, and +1.6% ATAC / +9.5% DNase profile Jensen–Shannon divergence versus ChromBPNet.
- For variant effect prediction, AlphaGenome matched or outperformed external models on 25 of 26 evaluations, including eQTL sign prediction +25.5% versus Borzoi and accessibility QTL +8.0% versus ChromBPNet. It achieved SOTA splicing variant prediction on six of seven benchmarks, with ClinVar auPRC 0.66 versus 0.64 by Pangolin for deep intronic/synonymous, 0.57 versus 0.55 for splice region, and 0.18 versus 0.16 for missense; on MFASS it reached auPRC 0.54 versus Pangolin 0.51 and SpliceAI/DeltaSplice 0.49.
- Multimodal scoring recapitulated TAL1 oncogene mechanisms: for chr. 1: 47239296: C>ACG, AlphaGenome predicted increased TAL1 expression 7.5 kb away, activating H3K27ac/H3K4me1, decreased repressive marks, and a MYB motif at the variant position.
Methods
- Model architecture: U-Net-inspired backbone producing 1D embeddings at 1-bp and 128-bp resolution and 2D embeddings at 2,048-bp resolution; convolutional layers model local patterns and transformer blocks model longer-range dependencies. Base-pair-resolution training on 1-Mb sequences uses sequence parallelism across eight interconnected TPU v3 devices.
- Training: two-stage process. Pretraining used a 4-fold cross-validation scheme to produce fold-specific models and all-fold teacher models; distillation then trained a single student model to reproduce an ensemble of all-fold teachers using augmented and mutationally perturbed input sequences. The distilled student runs in less than 1 s on an NVIDIA H100 GPU per variant.
- Evaluation: 24 genome track evaluations across all 11 modalities and 26 variant effect benchmarks, compared against the strongest available external model per task. Ablations assessed target resolution, sequence length, ensembling, distillation and multimodal training. Detailed methods are in the Supplementary Information.
Limitations
- Stated by the paper: accurately capturing distal regulatory elements more than 100 kb away remains a continuing objective; recapitulating tissue-specific patterns across cellular contexts and predicting condition-specific variant effects remain challenging; training and evaluations are heavily focused on protein-coding genes, with future work needed for non-coding genes such as microRNAs; species coverage is limited to human and mouse and evaluations are primarily human-focused; the model has not been benchmarked on personal genome prediction; and complex trait analysis is limited because AlphaGenome predicts molecular consequences rather than broader biological and environmental contributions.
- Also stated: performance decays with distance to the target gene/TSS and to the polyadenylation site; high score thresholds enrich for causal variant candidates but at low recall, particularly for GWAS variants; and further improvements are needed for intermediate splicing efficiencies and tissue-specific splicing nuances.
- Evident from the text: the paper reports occasional underprediction of expression impact for alternative polyadenylation and notes that both AlphaGenome and Borzoi still underestimate the impact of very distal enhancers.
Model-written summary. Check it against the paper before relying on it.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
94 Own result · 20 Own interpretation · 18 No basis given · 9 Sources not found · 5 Supported · 5 Partly supported · 4 Not checked yet · 2 Supported so far · 2 No full text of the sources · 1 Couldn’t confirm · 1 Own method · accuracy not yet measured
Every source the paper cites for this claim was checked and supports it.
In the paper
“Given that more than 98% of observed genetic variation in humans is non-coding7, global characterization of the complex effects of this vast majority of variants remains intractable without computational predictions.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Supportedcites [7] Halldorsson, B. V. et al. The sequences of 150,119 genomes in the UK Biobank. Nature 607, 732–740 (2022). source
Table 1 in “SNPs and indels” lists 6,380,795 coding variants among approximately 558 million WGS variants across its annotation categories. Coding variants therefore account for about 1.1% of those observed, supporting the claim that more than 98% are non-coding in this human cohort. (LLM, conf=0.88)
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“The ability of AlphaGenome to simultaneously score variant effects across all modalities accurately recapitulates the mechanisms of clinically relevant variants near the TAL1 oncogene6.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Abstract
What the sources say
- Partly supportedcites [6] Mansour, M. R. et al. Oncogene regulation. An oncogenic super-enhancer formed through somatic mutation of a noncoding intergenic element. Science 346, 1373–1377 (2014). sourceExperimental — accuracy not yet measured
The supplied passages (labeled Abstract in our copy) describe TAL1-region mutations in T-ALL samples and their mechanism: newly created MYB binding sites nucleate a super-enhancer that increases TAL1 expression. They do not evaluate AlphaGenome, its scoring across modalities, or whether its predictions accurately recapitulate those mechanisms. (LLM, conf=0.88)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Existing methods involve a trade-off between input sequence length and prediction resolution, thereby limiting their modality scope and performance1–5.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Abstract
What the sources say
- Partly supportedcites [1] Avsec, Ž. et al. Effective gene expression prediction from sequence by integrating long-range interactions. Nat. Methods 18, 1196–1203 (2021). sourceExperimental — accuracy not yet measured
The Methods (Model architecture; page unavailable) describe a trade-off for Enformer: finer prediction resolution would increase the sequence length entering its attention layers and make computation intractable on available hardware. The Results (Enformer improves gene expression prediction; page unavailable) also say previous models reached less-distant elements. The paper does not establish that existing methods generally trade input length against resolution or that this trade-off limits their modality scope. (LLM, conf=0.82)
- Partly supportedcites [2] Linder, J., Srivastava, D. & Yuan, H. Predicting RNA-seq coverage from DNA sequence as a unifying model of gene regulation. Nat. Genet. 57, 949–961 (2025). sourceExperimental — accuracy not yet measured
The Results (RNA-seq model design; page unavailable) explicitly describe a computational trade-off between sequence length and prediction resolution. The Methods (Model; page unavailable) explain how insufficient length or coarse resolution can obstruct learning, and the Main section describes earlier models’ narrower assay focus. The paper does not explicitly establish that this trade-off is what limits existing methods’ modality scope and performance overall. (LLM, conf=0.82)
- Partly supportedcites [3] Zhou, J. Sequence-based modeling of three-dimensional genome architecture from kilobase to chromosome scale. Nat. Genet. 54, 725–734 (2022). sourceExperimental — accuracy not yet measured
The Introduction (page unavailable) says earlier sequence models did not predict large-scale genome organization beyond 1 Mb, limiting predictions of large structures and structural-variant effects. The Results describe using coarser resolution at longer distances and report a small improvement in local predictions from larger sequence context. The paper does not establish that a length–resolution trade-off in existing methods limits their modality scope or overall performance. (LLM, conf=0.83)
- Not checked — no full text of the sourcecites [4] Jaganathan, K. et al. Predicting splicing from primary sequence with deep learning. Cell 176, 535–548 (2019). source
- Partly supportedcites [5] Kelley, D. R., Snoek, J. & Rinn, J. L. Basset: learning the regulatory code of the accessible genome with deep convolutional neural networks. Genome Res. 26, 990–999 (2016). sourceExperimental — accuracy not yet measured
The Discussion says prior sequence-kernel methods lose positional information, while position-specific kernels can suffer in predictive performance; it also says Basset offers finer nucleotide-level resolution than annotation-overlap approaches. It does not describe a trade-off between input sequence length and prediction resolution or say that such a trade-off limits modality scope. (LLM, conf=0.78)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“For instance, 99% (465 of 471) of validated enhancer–gene pairs fall within 1 Mb (ref. 12).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Unifying DNA sequence-to-function model
What the sources say
- Partly supportedcites [12] Gschwind, A. R. et al. An encyclopedia of enhancer-gene regulatory interactions in the human genome. Preprint at bioRxiv https://doi.org/10.1101/2023.11.09.563812 (2023). sourceExperimental — accuracy not yet measured
The Genome-wide maps of regulatory interactions section (page unavailable) says most predicted enhancer–gene interactions span short distances and that this is consistent with CRISPR perturbation results. The Supervised classification section reports 472 CRISPR-positive pairs, but our copy does not give the claimed count of 465 of 471 or establish that 99% of validated pairs fall within 1 Mb. (LLM, conf=0.83)
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
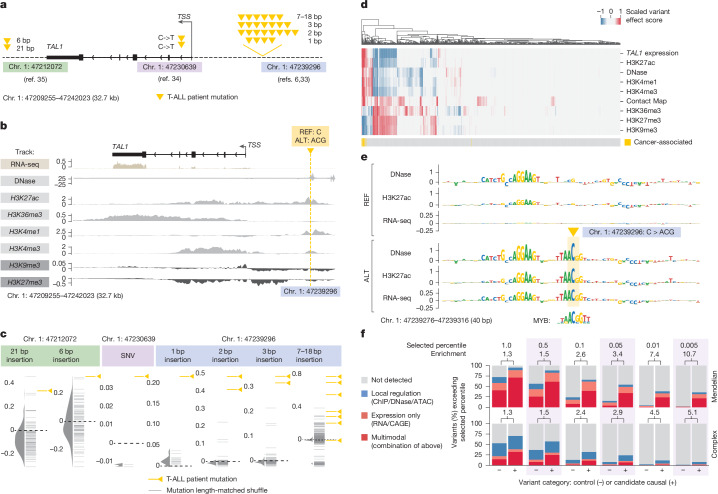
“Detailed ALT–REF predictions for an oncogenic insertion (chr. 1: 47239296: C>ACG) characterized in ref. 6. Shown are differences between AlphaGenome predictions between the ALT and REF sequences of the variant in CD34+ CMP tracks. The ALT sequence increases expression of the TAL1 gene 7.5 kb away.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Partly supportedcites [6] Mansour, M. R. et al. Oncogene regulation. An oncogenic super-enhancer formed through somatic mutation of a noncoding intergenic element. Science 346, 1373–1377 (2014). sourceExperimental — accuracy not yet measured
The paper’s main text (page unavailable in our copy) describes oncogenic insertions at a hotspot 7.5 kb upstream of TAL1 and concludes that they drive TAL1 overexpression. It does not identify the insertion as chr. 1:47239296:C>ACG, so our copy does not establish that this exact ALT allele is one it characterized. The AlphaGenome predictions are the citing authors’ analysis, not a finding attributed to this source. (LLM, conf=0.78)
plaindirectSingle case- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“We evaluated three groups of gain-of-function mutations affecting the TAL1 gene in T cell acute lymphoblastic leukaemia (T-ALL): a cluster of 5′ neo-enhancer mutations upstream of the TAL1 TSS6,33, an intronic single-nucleotide variant (SNV)34 and a 3′ neo-enhancer35.”
- Also stated in: § Multimodal view of variant effects · boosted
“All three variant groups were shown to converge on a common mechanism—upregulation of the TAL1 oncogene (Fig. 6a,b).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Multimodal view of variant effects
What the sources say
- Supportedcites [6] Mansour, M. R. et al. Oncogene regulation. An oncogenic super-enhancer formed through somatic mutation of a noncoding intergenic element. Science 346, 1373–1377 (2014). source
The citation to this paper is attached to the 5′ neo-enhancer mutations, not the separately cited intronic SNV or 3′ neo-enhancer. The passage describing a hotspot 7.5 kb upstream of the TAL1 transcription start site reports that its indel mutations generate a super-enhancer driving TAL1 overexpression in T-ALL. (LLM, conf=0.93)
- Supportedcites [33] Liu, Y. et al. The genomic landscape of pediatric and young adult T-lineage acute lymphoblastic leukemia. Nat. Genet. 49, 1211–1218 (2017). source
The section “Somatic alterations targeting transcriptional regulators” reports upstream TAL1 non-coding mutations that create a MYB-binding site and identifies 15 cases with identical or similar mutations. This supports the 5′ upstream mutation group to which this source is cited in the printed sentence; the intronic SNV and 3′ enhancer have separate citations. (LLM, conf=0.91)
- Supportedcites [34] Liu, Y. et al. Discovery of regulatory noncoding variants in individual cancer genomes by using cis-X. Nat. Genet. 52, 811–818 (2020). source
The citation is attached to the intronic SNV, not to all three mutation groups. In Results, “A recurrent intronic mutation activating TAL1 expression,” the paper identifies a recurrent C-to-T mutation in TAL1 intron 1 in T-ALL and links it to increased TAL1 expression and enhancer activity. (LLM, conf=0.93)
- Supportedcites [35] Smith, C. et al. TAL1 activation in T-cell acute lymphoblastic leukemia: a novel oncogenic 3′ neo-enhancer. Haematologica 108, 1259–1271 (2023). source
The Introduction describes upstream microinsertions, an intronic point mutation, and a newly discovered downstream microinsertion as distinct routes to TAL1 expression in T-ALL. The Abstract and Discussion report that the downstream mutation forms a 3′ neo-enhancer that activates TAL1. The paper supports convergence on TAL1 upregulation, though it does not say all three mutations use the same transcription factor. (LLM, conf=0.92)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Also stated in: § Multimodal view of variant effects · boosted
The sources with a verdict support this claim; the rest weren’t checked yet, have no full text we could read, or their verdicts aren’t rated yet.
In the paper
“We have not yet benchmarked the model on personal genome prediction, which is a known weakness of models in this space40,41.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Discussion
What the sources say
- Not checked yetcites [40] Sasse, A. et al. Benchmarking of deep neural networks for predicting personal gene expression from DNA sequence highlights shortcomings. Nat. Genet. 55, 2060–2064 (2023). source
Checked before — its earlier verdict is held back until a new check runs.
- Supportedcites [41] Huang, C. et al. Personal transcriptome variation is poorly explained by current genomic deep learning models. Nat. Genet. 55, 2056–2059 (2023). source
The citation supports the claim that personal-genome prediction is a weakness of models in this space, not the citing authors’ statement that they have not yet benchmarked AlphaGenome. The Main section (page not supplied) reports that four sequence-to-expression models underperform when predicting expression differences across individuals from personal DNA sequences. (LLM, conf=0.92)
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Not checked yet
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“Here we present AlphaGenome, a model that unifies multimodal prediction, long-sequence context and base-pair resolution into a single framework.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“The modalities include gene expression, transcription initiation, chromatin accessibility, histone modifications, transcription factor binding, chromatin contact maps, splice site usage and splice junction coordinates and strength.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Abstract
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“The splicing predictions of AlphaGenome include a new splice junction prediction approach alongside splice site usage prediction.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- It may state two separate claims as one. 44
- Probably not a substantial claim. 27
- The quote only partly states this claim. 1
Every source the paper cites for this claim was checked and supports it.
In the paper
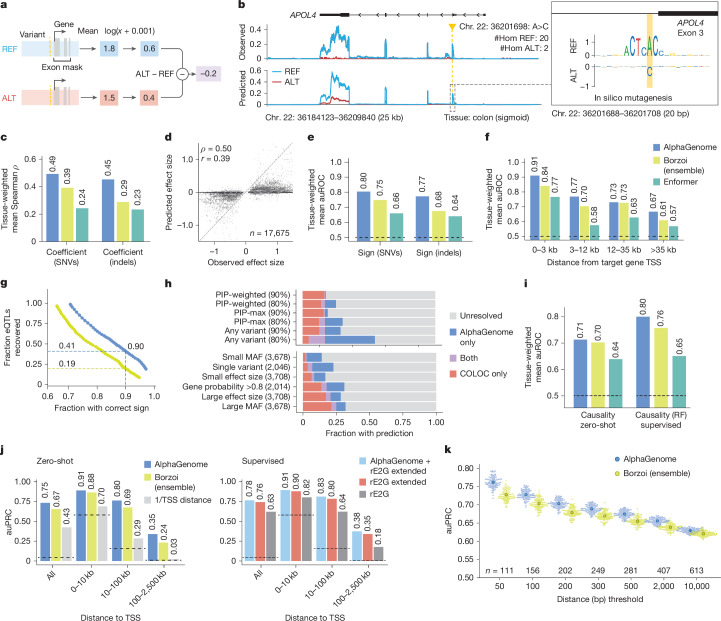
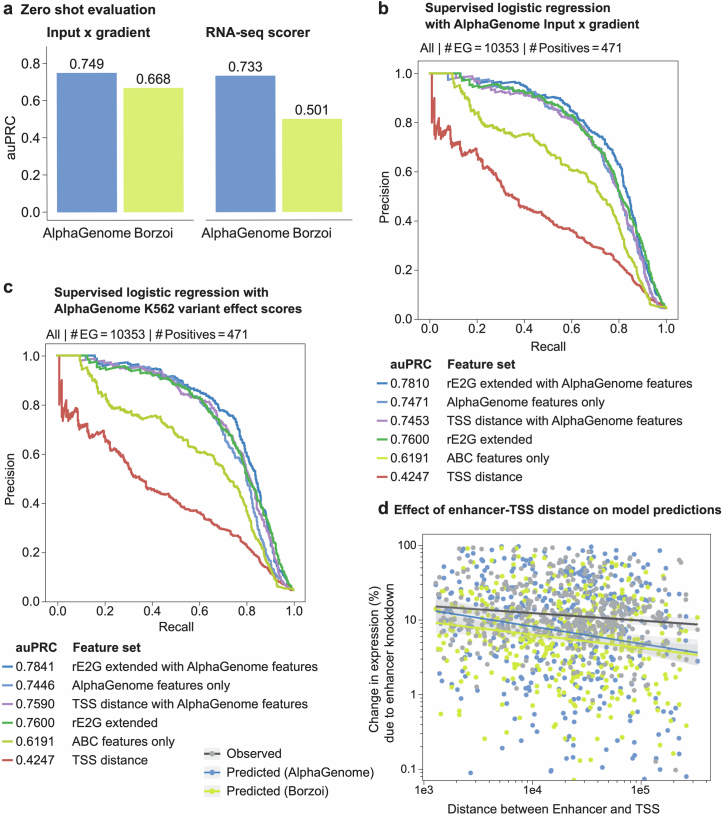
“Enhancer–gene linking performance (ENCODE–rE2G CRISPRi dataset12). Zero-shot evaluation: performance (auPRC) comparison stratified by enhancer-to-TSS distance. Supervised evaluation: AlphaGenome input gradient score integrated into ENCODE–rE2G extended and ENCODE–rE2G models.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Supportedcites [12] Gschwind, A. R. et al. An encyclopedia of enhancer-gene regulatory interactions in the human genome. Preprint at bioRxiv https://doi.org/10.1101/2023.11.09.563812 (2023). source
The citation is attached to the identification of the ENCODE–rE2G CRISPRi dataset, not to the citing paper’s AlphaGenome evaluations. In “Supervised classification of enhancer-gene regulatory interactions” (page unavailable), the source describes its aggregated CRISPR-tested element–gene dataset, drawn mostly from CRISPRi experiments, and its use to train and evaluate ENCODE-rE2G. (LLM, conf=0.91)
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“This distilled student model, as shown previously14, achieved improved robustness and variant effect prediction accuracy in a single model instance, making predictions across all modelled modalities and cell types with a single device call per variant.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Unifying DNA sequence-to-function model
What the sources say
- Supportedcites [14] Zhou, J., Rizzo, K., Tang, Z. & Koo, P. K. Uncertainty-aware genomic deep learning with knowledge distillation. Preprint at bioRxiv https://doi.org/10.1101/2024.11.13.623485 (2024). source
The Results section on zero-shot variant effect generalization reports that distilled models generally had stronger correlations with experimentally measured variant effects than standard-trained models. The Results section on covariate shifts reports improved generalization with augmentation, and the section on approximating deep ensembles identifies the student as an individual model trained from an ensemble. (LLM, conf=0.91)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
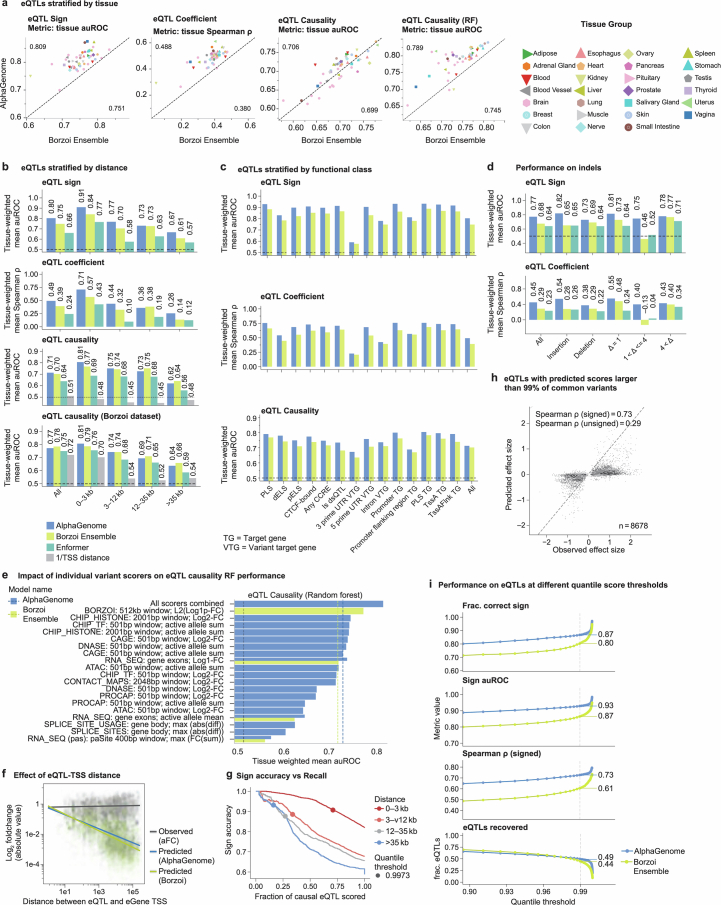
“As previously reported26, performance decays with distance to the target gene across all eQTL tasks (Extended Data Fig. 5f).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of eQTL effects
What the sources say
- Partly supportedcites [26] Karollus, A., Mauermeier, T. & Gagneur, J. Current sequence-based models capture gene expression determinants in promoters but mostly ignore distal enhancers. Genome Biol. 24, 56 (2023). sourceExperimental — accuracy not yet measured
In Results, ‘Most distal eQTLs do not have a meaningful impact on expression predictions’ (Fig. 6A–B), the paper reports that Enformer’s predicted eQTL effects decay rapidly with distance to the target gene’s TSS. It does not establish that performance decays across all eQTL tasks; it also reports no systematic distance-related decay in the measured eQTL effect sizes. (LLM, conf=0.86)
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
- Partly supported
The sources with a verdict support this claim; the rest weren’t checked yet, have no full text we could read, or their verdicts aren’t rated yet.
In the paper
“Beyond impacting isoform composition through splicing modulation, non-coding variants can influence traits and cause diseases by altering gene expression23,24.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks
What the sources say
- Not checked — no full text of the sourcecites [23] Albert, F. W. & Kruglyak, L. The role of regulatory variation in complex traits and disease. Nat. Rev. Genet. 16, 197–212 (2015). source
- Supportedcites [24] Maurano, M. T. et al. Systematic localization of common disease-associated variation in regulatory DNA. Science 337, 1190–1195 (2012). source
The Abstract links disease- and trait-associated noncoding variants to transcriptional regulatory mechanisms, including promoters, enhancers, and expression quantitative trait loci. The Discussion reports that disease- and trait-associated variants occur in regulatory DNA and can perturb transcription factor networks, supporting the claim that altered gene regulation can influence traits and disease. No page numbers are provided in our copy. (LLM, conf=0.89)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. It may state two separate claims as one.
The cited sources that could be checked couldn’t confirm or dispute this claim.
In the paper
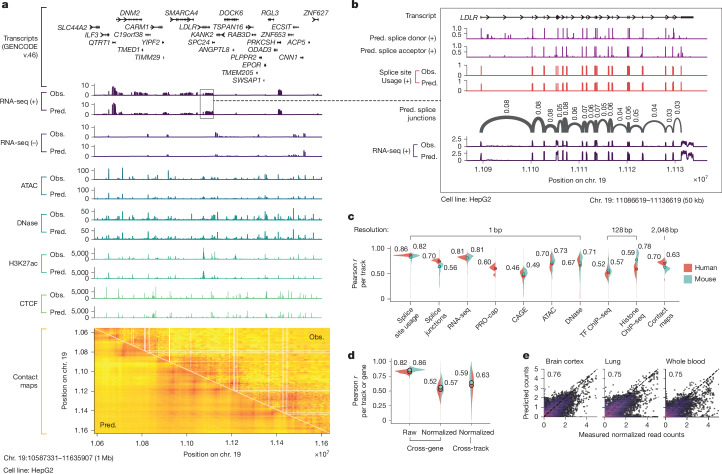
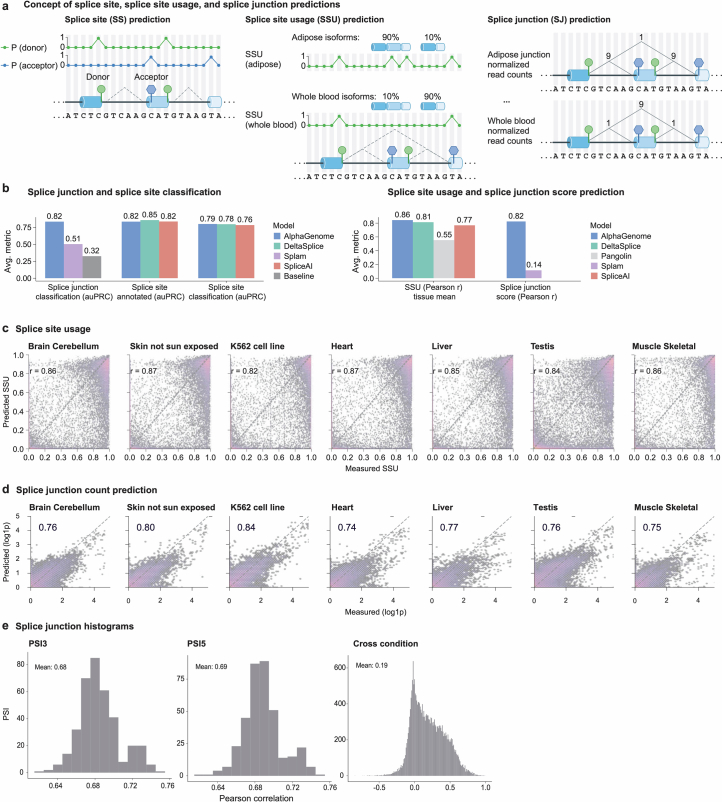
“Predicted versus observed splice junction read counts (log(1 + x) transformed; n = 1,344,738) and Pearson r between them in selected human tissues known for having distinct splicing patterns49.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Could not verifycites [49] Ling, J. P. et al. ASCOT identifies key regulators of neuronal subtype-specific splicing. Nat. Commun. 11, 137 (2020). sourceExperimental — accuracy not yet measured
The Methods section on Snaptron compilations describes tables of observed splice-junction counts, and the Results discuss splicing patterns in human tissues. Neither passage reports a predicted-versus-observed count comparison, a log(1 + x) transformation, Pearson r, or n = 1,344,738. Searches for those terms found no matching passage in our copy. (LLM, conf=0.72)
plaincorrelationaln = 1,344,738- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. It may state two separate claims as one.
- Could not verify
Every source the paper cites for this claim was checked and supports it.
In the paper
“By contrast, the alternative sequence introduced a MYB motif at the variant position predicted to increase TAL1 expression, chromatin accessibility and H3K27ac chromatin marks (Fig. 6e), as discovered previously6.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Multimodal view of variant effects
What the sources say
- Supportedcites [6] Mansour, M. R. et al. Oncogene regulation. An oncogenic super-enhancer formed through somatic mutation of a noncoding intergenic element. Science 346, 1373–1377 (2014). source
The supplied passages (labelled Abstract) report that mutations at the TAL1 enhancer introduce MYB binding motifs and drive TAL1 overexpression. They also propose that MYB recruits CBP, producing abundant H3K27 acetylation that opens chromatin; this supports the citing sentence’s predicted increases in H3K27ac and accessibility. (LLM, conf=0.91)
plaindirectSingle case- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“The model takes 1 Mb of DNA sequence as input and predicts a diverse range of genome tracks across numerous cell types.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“AlphaGenome achieved SOTA performance on 22 of 24 genome track prediction tasks and 25 of 26 variant effect prediction tasks.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“On splicing (Extended Data Fig. 2a), AlphaGenome accurately predicts splice sites (Extended Data Fig. 2b) and splice site usage (Extended Data Fig. 2b,c).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved track prediction performance
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
None of the sources cited for this claim have been checked yet.
In the paper
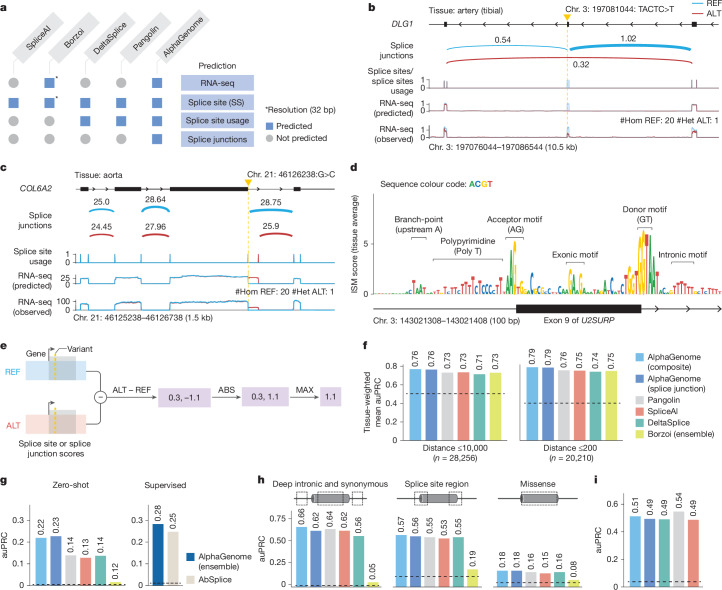
“MFASS splicing variant classification (MPRA-tested variants). auPRC on the classification of experimentally validated splice-disrupting variants (data from Chong et al.22).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Not checked yetcites Chong et al. [22] Chong, R. et al. A multiplexed assay for exon recognition reveals that an unappreciated fraction of rare genetic variants cause large-effect splicing disruptions. Mol. Cell 73, 183–194 (2019). source
Checked before — its earlier verdict is held back until a new check runs.
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
- Not checked yet
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Although AlphaGenome demonstrates the ability to predict tissue-specific alternative splicing in some instances (Supplementary Fig. 1), further improvements are needed to precisely predict intermediate splicing efficiencies and to capture tissue-specific nuances (Extended Data Fig. 2c,e).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved track prediction performance
boosted- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“AlphaGenome demonstrated improved prediction of both the magnitude (‘coefficient’; Spearman ρ with SuSiE25 β posterior) and direction (‘sign’; area under the receiver operating characteristic (auROC) curve) of eQTL effects compared with the previous SOTA (Borzoi) (Fig. 4c–f).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of eQTL effects
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
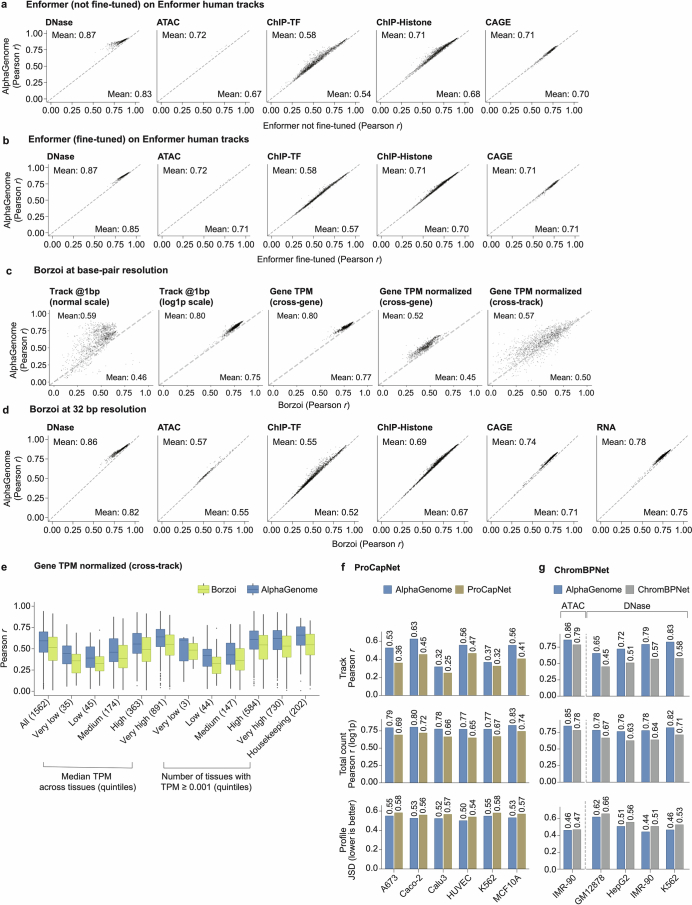
“Track prediction performance evaluation across different modalities. Violin plots display the distribution of Pearson correlations between predicted and observed tracks evaluated on held-out test intervals.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“its composite scores outperformed the existing best method in each category across all three variant categories: deep intronic and synonymous (area under the precision–recall curve (auPRC) 0.66 versus 0.64 by Pangolin), splice region (auPRC 0.57 versus 0.55 by Pangolin) and missense (auPRC 0.18 versus 0.16 by DeltaSplice and Pangolin”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved splicing variant predictions
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
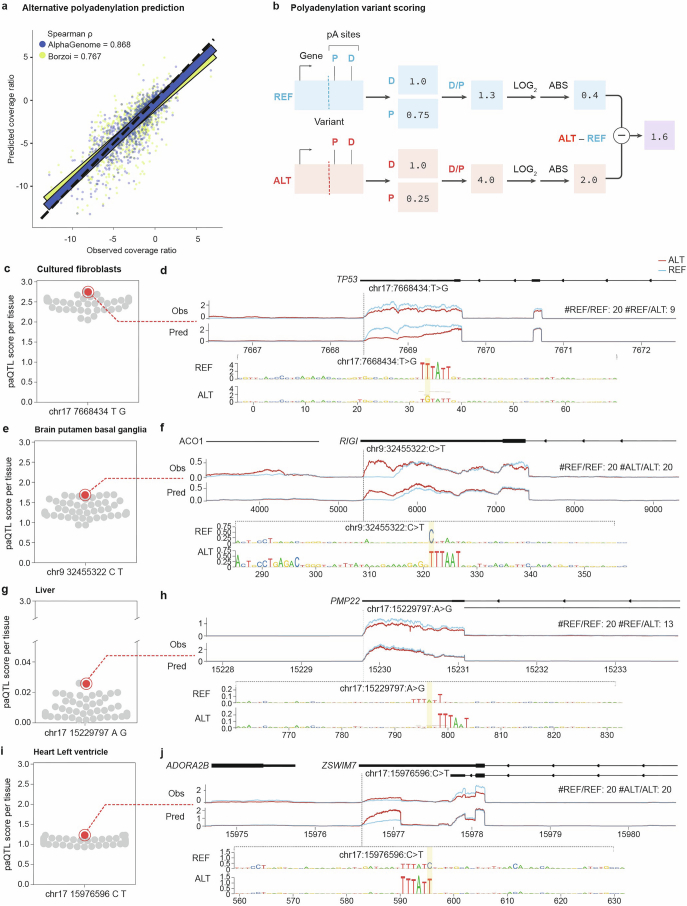
“Performance (auPRC) of paQTL variant effect prediction, thresholded by distance from the polyadenylation site.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“AlphaGenome predicts all three of these quantities alongside direct RNA-seq coverage prediction, thereby providing a more comprehensive view of the splicing-related molecular consequences of variants (Fig. 3a).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved splicing variant predictions
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
In the paper
“Coverage of predictions across GWAS loci. Fraction of GWAS credible sets (from Open Targets52) with a predicted direction of effect for a plausible target gene, comparing AlphaGenome predictions to the eQTL co-localization approach.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Not matchedcites Open Targets52
Couldn’t be matched to a paper in Crossref, so it can’t be checked.
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
- Not matched
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
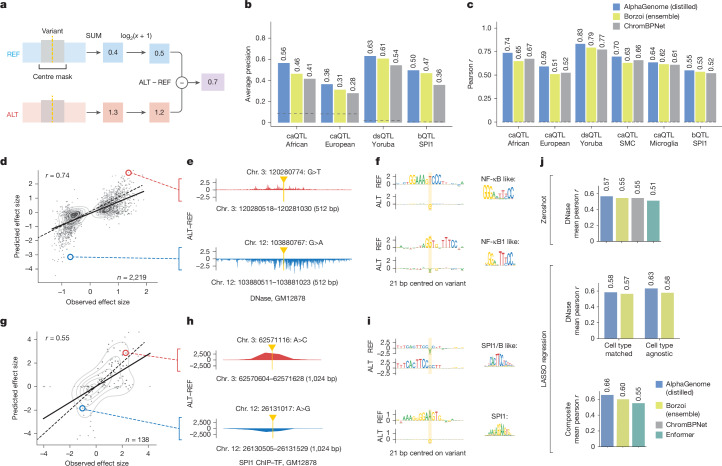
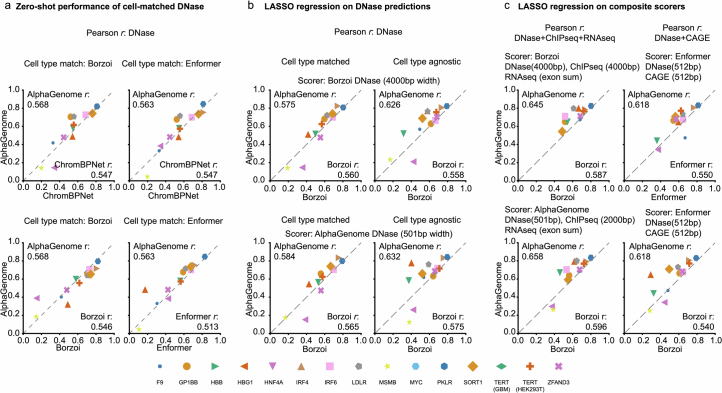
“CAGI5 MPRA challenge performance (average across loci; mean Pearson r). Top, zero-shot using cell-type-matched DNase; middle, LASSO regression using cell-type-matched or agnostic DNase; bottom, LASSO regression using multimodal features (DNase + RNA + histone ChIP-seq output types for AlphaGenome and Borzoi; DNase + CAGE output types for Enformer) and all cell types.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“eQTL sign prediction performance stratified by different variant-to-TSS distance bins (SNVs only).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Performance comparison of AlphaGenome, Borzoi and ChromBPNet on QTL causality (b; average precision) and QTL effect size (c; Pearson r) across QTL types and ancestries.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Comparison of performance (auROC) at distinguishing causal from non-causal eQTLs (‘causality’) using both zero-shot and supervised approaches (Methods).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Comparison of performance (Spearman’s ρ) at predicting the effect size of eQTLs across 49 GTEx tissues (‘coefficient’) for different models and variant sets.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Although overall expression levels are predicted well, accurately capturing cell-type-specific expression deviations remains a challenging task (Fig. 2d and Supplementary Fig. 2j).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved track prediction performance
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
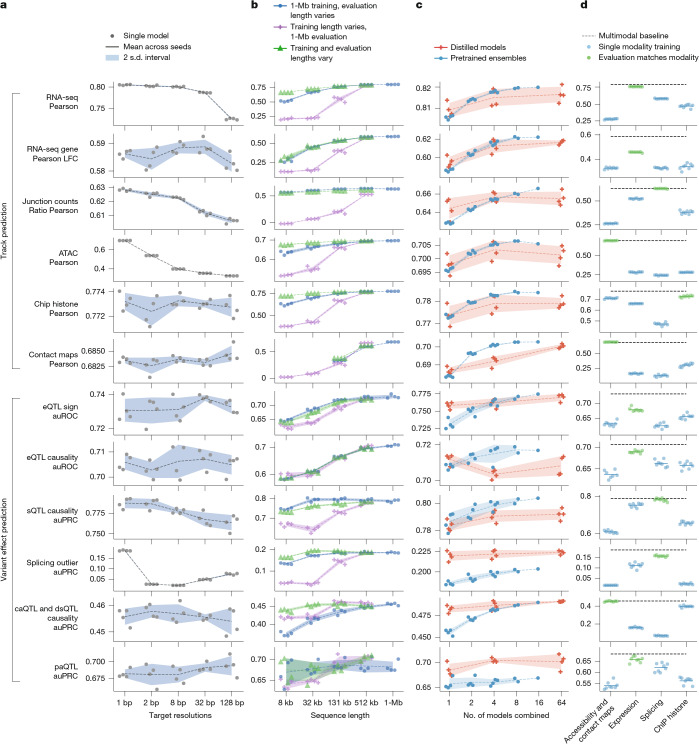
“Impact of the number of sub-models in ensembling and distillation. Performance comparison for mean ensembles of pretrained models (blue dots/contours; x axis indicates ensemble size) versus single models produced by distillation using 1, 4 or 64 teacher models (orange crosses/contours; x axis indicates number of teachers).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelationaln = 4- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Using a threshold calibrated to 80% accuracy on eQTLs (Fig. 4g), AlphaGenome assigned a confident sign prediction for at least one variant in 49% of GWAS credible sets (11% using a conservative PIP-weighted scoring approach; Fig. 4h).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of eQTL effects
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“The alternative ‘C’ allele is associated with lower expression of APOL4 in GTEx data (sum of single effects (SuSiE) ‘β posterior’ = −0.709; posterior inclusion probability (PIP) > 0.9), and both the RNA-seq coverage and direction of effect are recapitulated by the predictions of AlphaGenome (variant score = −1.52; quantile score = −1.00; Methods).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of eQTL effects
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“AlphaGenome improved the tissue-weighted mean Spearman ρ from 0.39 to 0.49 and the mean sign auROC from 0.75 to 0.80.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of eQTL effects
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“RNA-seq, ATAC-seq and DNase-seq track predictions are at 1-bp resolution; H3K27ac and CTCF ChIP-seq are at 128-bp resolution; and contact maps are at 2,048-bp resolution.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it. It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Impact of multimodal learning. Performance comparison evaluating models trained only on specific modality groups (blue dots; n = 8 seeds per group, highlighted in green if the modality matches the evaluation metric) against the full multimodal model (black dashed line; n = 4 seeds average).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plainn = 8- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Comparison of performance (auROC) at predicting the direction of effect of eQTLs (‘sign’) for different models and variant sets.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
In the paper
“Comparison of AlphaGenome composite and splice junction scorers versus other methods for classifying fine-mapped sQTL variants. Variants are stratified into two groups by distance to the splice site, as done in Borzoi2. Tissue-specific auPRCs were averaged and weighted by variant count per tissue.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Not matchedcites Borzoi2
Couldn’t be matched to a paper in Crossref, so it can’t be checked.
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. It may state two separate claims as one.
- Not matched
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Predicted TAL1 expression change (ALT–REF) in CD34+ CMPs. RNA-seq variant scores for TAL1 expression in CD34+ CMPs. Oncogenic mutations (orange) are compared with randomly sampled, length-matched indels (grey).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
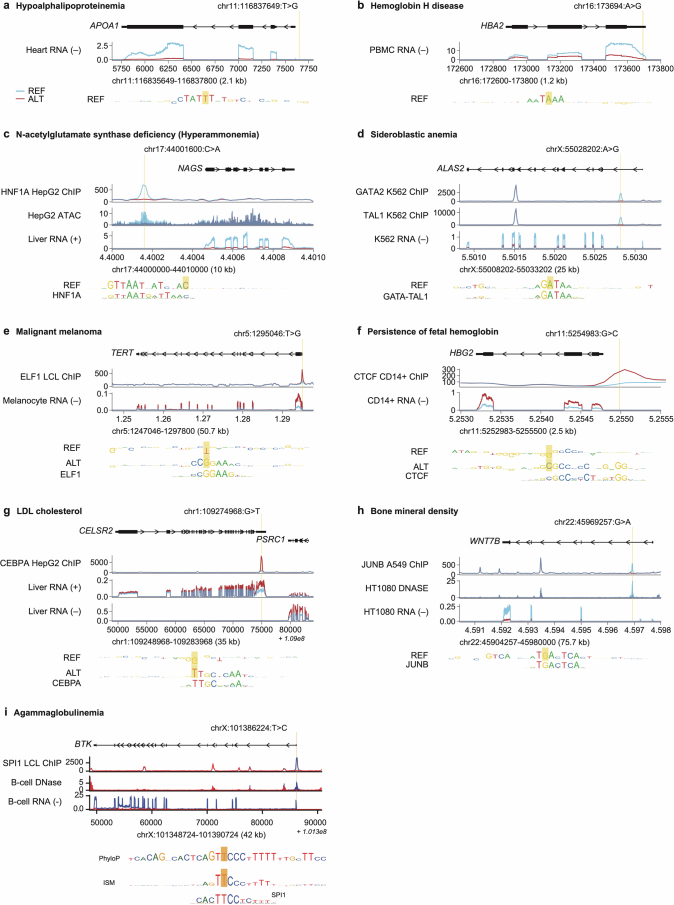
“AlphaGenome accurately predicts variant effects on chromatin accessibility and SPI1 transcription factor binding.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“Taking less than 1 s on an NVIDIA H100 GPU, the student model is highly efficient for large-scale variant effect prediction relative to the alternative approach of ensembling several independently trained models.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Unifying DNA sequence-to-function model
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“AlphaGenome accurately predicted this established consequence across all levels: a substantial reduction in the predicted usage of the affected exon’s splice site, loss of predicted junctions linking the skipped exon edges, emergence of a putative junction bypassing the exon and strong decrease in predicted RNA-seq coverage of the exon.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved splicing variant predictions
plaindirectSingle case- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Notably, AlphaGenome exhibited a +14.7% relative improvement in cell-type-specific gene-level expression log-fold change prediction compared with Borzoi2, another multimodal sequence model (Fig. 1e and stratified metrics in Extended Data Fig. 3e).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance overview
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
The quote only partly states this claim.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Impact of target resolution. Performance comparison across models trained to predict targets (DNA accessibility, gene expression and splicing) at varying resolutions (x axis; 1–128 bp).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelationaln = 4- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Comparison of AlphaGenome-predicted variant scores and observed effect sizes (SuSiE β posterior) for 17,675 fine-mapped GTEx eQTLs (SNVs). Each point is a unique variant/gene/tissue combination. Spearman’s ρ (signed) = 0.50; Spearman’s ρ (unsigned; absolute values) = 0.10. Pearson’s r (signed) = 0.39; Pearson’s r (unsigned; absolute values) = 0.20.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“When assessing the ability of AlphaGenome to predict whether rare variants disrupt splicing using data from a massively parallel splicing minigene reporter assay (multiplexed functional assay of splicing using SORT-seq (MFASS))22, it was outperformed by Pangolin (auPRC 0.54 versus 0.51) but surpassed SpliceAI and DeltaSplice (both auPRC 0.49; Fig. 3i).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved splicing variant predictions
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Setting a high threshold on the quantitative scores of AlphaGenome (for example, predicted expression change) strongly enriched for causal variant candidates among matched controls (Fig. 6f), although at the cost of low recall, particularly for GWAS variants.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Multimodal view of variant effects
boostedcorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Cumulative addition experiments showed that although track predictions improved most when their relevant data types were added, variant effect tasks benefited most from the initial inclusion of expression and accessibility data, with diminishing returns from subsequent modality additions (Supplementary Fig. 12, right panel).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Model and data ablations
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“This benefit was task-dependent; for example, predicting accessibility variants was effective using only accessibility data, whereas predicting eQTLs benefited from the full multimodal input (a more complete set of experiments is shown in Supplementary Fig. 12).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Model and data ablations
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“ISM revealed that AlphaGenome learned the relevance of the canonical polyadenylation motif and could detect variants that disrupt or create this motif (Extended Data Fig. 6c–h), although it occasionally underpredicted the expression impact (Extended Data Fig. 6i,j).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of 3′ polyadenylation QTL effects
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“This strong performance generalized across more datasets, including European ancestry caQTLs, Yoruba ancestry DNase sensitivity QTLs and predictions within specific cell types such as microglia and cardiac smooth muscle cells.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved prediction of chromatin accessibility, DNase sensitivity and binding QTLs
boostedcorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“However, leveraging the predictions of AlphaGenome within a supervised framework boosted performance on the causality task; training a random forest model using the scores of AlphaGenome from several modalities increased the mean auROC from 0.68 to 0.75, also surpassing previous SOTA performance (mean auROC 0.71 for Borzoi; Fig. 4i).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of eQTL effects
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“It also predicted decreased levels of the repressive histone marks H3K9me3 and H3K27me3 near the TAL1 TSS and elevated active transcription mark H3K36me3 across the TAL1 gene body, both of which are concordant with its predicted increase in TAL1 mRNA expression levels (Fig. 6b).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Multimodal view of variant effects
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Rests on an assumption: CD34+ common myeloid progenitor (CMP) data is the closest available match to the T-ALL cell of origin.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“It also strongly outperformed the simpler DNase-based ENCODE–rE2G model and a distance-to-TSS baseline.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Competitive enhancer–gene linking
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Complementary experiments showed that excluding single modality groups during training typically resulted in only modest performance decreases, suggesting redundancy between modalities (Supplementary Fig. 12, left panel).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Model and data ablations
hedgeddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“AlphaGenome consistently achieved SOTA performance compared with both Borzoi and the specialized ChromBPNet model. This was observed across QTL types and ancestries, both in predicting QTL causality (average precision; Fig. 5b and Supplementary Fig. 9) and in correlating with experimentally determined effect sizes (‘coefficient’; Pearson r; Fig. 5c and Supplementary Fig. 9).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved prediction of chromatin accessibility, DNase sensitivity and binding QTLs
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A result the paper reports from its own data — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“AlphaGenome outperformed Borzoi at all considered distances, although the predictive accuracy of both models declined with distance to the polyadenylation site (Fig. 4k and Supplementary Table 5).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of 3′ polyadenylation QTL effects
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“Our species coverage remains limited to human and mouse, and the evaluations in this study are primarily human-focused.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Discussion
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
None of the sources cited for this claim could be matched to a paper in Crossref, so there is nothing to check it against.
In the paper
“Models such as Enformer1 and Borzoi2 can process longer sequences (approximately 200–500 kb) to capture broader context but at the cost of reducing output resolution (128-bp or 32-bp bins), which can blur fine-scale regulatory features such as splice sites, transcription factor footprints or polyadenylation sites.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Not matchedcites Enformer1
Couldn’t be matched to a paper in Crossref, so it can’t be checked.
- Not matchedcites Borzoi2
Couldn’t be matched to a paper in Crossref, so it can’t be checked.
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Not matched
None of the sources cited for this claim could be matched to a paper in Crossref, so there is nothing to check it against.
In the paper
“Although models such as SpliceAI4, BPNet8 and ProCapNet9 provide base-resolution predictions, they are restricted to short input sequences (for example, 10 kb or less), and thus may miss the influence of distal regulatory elements.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
What the sources say
- Not matchedcites SpliceAI4
Couldn’t be matched to a paper in Crossref, so it can’t be checked.
- Not matchedcites BPNet8
Couldn’t be matched to a paper in Crossref, so it can’t be checked.
- Not matchedcites ProCapNet9
Couldn’t be matched to a paper in Crossref, so it can’t be checked.
hedgedconditional- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Not matched
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“A second trade-off exists between capturing diverse modalities versus specializing in one or a few.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“However, these more generalist models can lag behind their specialized counterparts on certain tasks, such as splicing, or may lack particular modalities, such as contact maps.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
hedgedconditional- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Furthermore, ISM around the variant indicates a possible nucleotide splice donor sequence motif, which is disrupted by the variant, thereby leading to an aberrant transcript and reduced expression (Fig. 4b; inset).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance across gene expression tasks > Improved prediction of eQTL effects
hedgeddirectSingle case- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
What the paper did — it cites no source for this, so there is none to check. The quote shows it is in the paper.
In the paper
“Both competing models and AlphaGenome pretrained fold-split models were evaluated on held-out genome regions unseen during model training.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
In the paper
“One of the main ways genetic variants cause disease is by disrupting splicing15, a process that produces mature RNA sequences by excising introns and ligating exons at splice junctions.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved splicing variant predictions
What the sources say
- Not checked — no full text of the sourcecites [15] López-Bigas, N., Audit, B., Ouzounis, C., Parra, G. & Guigó, R. Are splicing mutations the most frequent cause of hereditary disease? FEBS Lett. 579, 1900–1903 (2005). source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Collectively, these results demonstrate that AlphaGenome more accurately models both genome tracks and variant effects.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance overview
boostedcorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Rests on an assumption: Generalization to unseen genome intervals is a prerequisite for high-quality variant effect prediction.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Moreover, although the model predicts tissue-specific and cell-type-specific genome tracks with some success, accurately recapitulating tissue-specific patterns across cellular contexts and predicting condition-specific variant effects remain challenging.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Discussion
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Furthermore, the new capability of AlphaGenome to directly model splice junctions enables a more holistic view of splice-disrupting variants.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Discussion
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: fails — The paper may give no reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it. The paper may give no reason why it holds.
None of the sources cited for this claim have been checked yet.
In the paper
“Although models of molecular effect do not directly predict phenotypic consequences (Supplementary Note (trait-altering variants) and Supplementary Fig. 11)”
§ Advancing regulatory variant effect prediction with AlphaGenome > Multimodal view of variant effects
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Accurately capturing the influence of distal regulatory elements (more than 100 kb away) remains a continuing objective.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Discussion
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Overall, these analyses highlight the strong performance of AlphaGenome in predicting variant effects on chromatin states in QTL benchmarks, as well as its ability to leverage accessibility predictions to model the regulatory activity of sequences as measured by MPRAs.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Improved prediction of chromatin accessibility, DNase sensitivity and binding QTLs
boostedcorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“In summary, AlphaGenome provides a powerful and unified model for analysing the regulatory genome.”
§ Advancing regulatory variant effect prediction with AlphaGenome > Discussion
boosted- Asserted: passes — The authors put it forward as true.
- Checkable: fails — It may be too vague to check against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: fails — The paper may give no reason why it holds.
Probably not a substantial claim. It may be too vague to check against evidence. The paper may give no reason why it holds.
The paper relies on this without support.
Successfully trained sequence-to-function models can accurately predict experimental measurements from input sequences.
“Successfully trained sequence-to-function models accurately predict experimental measurements from input sequences.”
The paper relies on this without support.
Comparing genome track predictions from an alternative sequence versus a reference sequence allows prediction of the molecular effects of variants.
“Furthermore, by comparing genome track predictions from an alternative sequence versus a reference sequence, these models can predict the molecular effects of variants.”
The paper relies on this without support.
Generalization to unseen genome intervals is a prerequisite for high-quality variant effect prediction.
“we first assessed its generalization to unseen genome intervals, a prerequisite for high-quality variant effect prediction.”
The paper relies on this without support.
Tissue-specific gene expression is modulated by enhancer–promoter interactions, often involving enhancers in distal genomic regions.
“given that tissue-specific gene expression is modulated by enhancer–promoter interactions, often involving enhancers in distal genomic regions.”
The paper relies on this without support.
CD34+ common myeloid progenitor (CMP) data is the closest available match to the T-ALL cell of origin.
“we analysed predictions in CD34+ common myeloid progenitor (CMP) data, the closest available match to the T-ALL cell of origin.”
The paper relies on this without support.
MPRAs measure the regulatory activity of short DNA sequences, a process closely linked to local chromatin accessibility and transcription factor binding.
“MPRAs measure the regulatory activity of short DNA sequences (typically through reporter gene expression), a process closely linked to local chromatin accessibility and transcription factor binding.”
The paper relies on this without support.
The RNA-seq tracks labelled thymus, CMP and haematopoietic multipotent progenitor cells are relevant to T-ALL biology.
“the difference in oncogenic versus shuffled mutations is most pronounced in T-ALL-relevant tracks such as ‘thymus’, ‘CMP’ and ‘haematopoietic multipotent progenitor cells’ (Supplementary Fig. 10).”
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
Each contradiction between two passages listed here was confirmed by an independent check.
Rated minor: it would not touch a result — a value no result rests on, a p-value that stays on the same side of the significance level, or a conflict that is clerical, resolved elsewhere in the paper, or outside its own findings.
In the paper
§ Advancing regulatory variant effect prediction with AlphaGenome > Performance overview
“Notably, AlphaGenome exhibited a +14.7% relative improvement in cell-type-specific gene-level expression log-fold change prediction compared with Borzoi2, another multimodal sequence model (Fig. 1e and stratified metrics in Extended Data Fig. 3e).”
§ Advancing regulatory variant effect prediction with AlphaGenome > Main
“e, Variant effect prediction: distilled all-fold model.”
Citations
Cites
U-Net: Convolutional Networks for Biomedical Image Segmentation
Ronneberger, O., Fischer, P., & Brox, T. (2015) · Lecture notes in computer science · doi:10.1007/978-3-319-24574-4_28 · Cited by 99,404
Scikit-learn: Machine Learning in Python
Pedregosa, F., Varoquaux, G., Gramfort, A., Michel, V., Thirion, B., Grisel, O., Blondel, M., Müller, A., Nothman, J., Louppe, G., et al. (2012) · arXiv · doi:10.48550/arxiv.1201.0490 · Cited by 71,016
STAR: ultrafast universal RNA-seq aligner
Dobin, A., Davis, C. A., Schlesinger, F., Drenkow, J., Zaleski, C., Jha, S., Batut, P., Chaisson, M., & Gingeras, T. R. (2013) · Bioinformatics · doi:10.1093/bioinformatics/bts635 · Cited by 51,709
Highly accurate protein structure prediction with AlphaFold
Jumper, J., Evans, R., Pritzel, A., Green, T., Figurnov, M., Ronneberger, O., Tunyasuvunakool, K., Bates, R., Žídek, A., Potapenko, A., et al. (2021) · Nature · doi:10.1038/s41586-021-03819-2 · Cited by 43,764
Self-citationAn integrated encyclopedia of DNA elements in the human genome
The ENCODE Project Consortium (2012) · Nature · doi:10.1038/nature11247 · Cited by 18,072
Twelve years of SAMtools and BCFtools
Danecek, P., Bonfield, J., Liddle, J., Marshall, J., Ohan, V., Pollard, M., Whitwham, A., Keane, T., McCarthy, S., Davies, R. M., et al. (2021) · GigaScience · doi:10.1093/gigascience/giab008 · Cited by 16,298
The Ensembl Variant Effect Predictor
McLaren, W., Gil, L., Hunt, S. E., Riat, H. S., Ritchie, G. R. S., Thormann, A., Flicek, P., & Cunningham, F. (2016) · Genome Biology · doi:10.1186/s13059-016-0974-4 · Cited by 8,099
Integrative analysis of 111 reference human epigenomes
Roadmap Epigenomics Consortium, Kundaje, A., Meuleman, W., Ernst, J., Bilenky, M., Yen, A., Heravi-Moussavi, A., Kheradpour, P., Zhang, Z., Wang, J., et al. (2015) · Nature · doi:10.1038/nature14248 · Cited by 6,368
The GTEx Consortium atlas of genetic regulatory effects across human tissues
The GTEx Consortium, Aguet, F., Anand, S., Ardlie, K. G., Gabriel, S., Getz, G. A., Graubert, A., Hadley, K., Handsaker, R. E., Huang, K. H., et al. (2020) · Science · doi:10.1126/science.aaz1776 · Cited by 5,547
Self-citationGiambartolomei, C., Vukcevic, D., Schadt, E. E., Franke, L., Hingorani, A. D., Wallace, C., & Plagnol, V. (2014) · PLoS Genetics · doi:10.1371/journal.pgen.1004383 · Cited by 4,261
ClinVar: improving access to variant interpretations and supporting evidence
Landrum, M. J., Lee, J. M., Benson, M., Brown, G. R., Chao, C., Chitipiralla, S., Gu, B., Hart, J., Hoffman, D., Jang, W., et al. (2018) · Nucleic Acids Research · doi:10.1093/nar/gkx1153 · Cited by 4,196
Systematic Localization of Common Disease-Associated Variation in Regulatory DNA
Maurano, M. T., Humbert, R., Rynes, E., Thurman, R. E., Haugen, E., Wang, H., Reynolds, A. P., Sandstrom, R., Qu, H., Brody, J., et al. (2012) · Science · doi:10.1126/science.1222794 · Cited by 3,716 · cited 1×
“Beyond impacting isoform composition through splicing modulation, non-coding variants can influence traits and cause diseases by altering gene expression 23,24 .”
Transformer-XL: Attentive language models beyond a fixed-length context
Dai, Z., Yang, Z., Yang, Y., Carbonell, J., Le, Q., & Salakhutdinov, R. (2019) · Proceedings of the 57th Annual Meeting of the Association for Computational Linguistics · doi:10.18653/v1/p19-1285 · Cited by 3,169
Predicting splicing from primary sequence with deep learning
Jaganathan, K., Kyriazopoulou Panagiotopoulou, S., McRae, J. F., Darbandi, S. F., Knowles, D., Li, Y. I., Kosmicki, J. A., Arbelaez, J., Cui, W., Schwartz, G. B., et al. (2019) · Cell · doi:10.1016/j.cell.2018.12.015 · Cited by 2,853 · cited 2×
“…outcomes can be modelled at three levels: the probability that any given nucleotide acts as a splice donor or acceptor (splice site prediction) 4,11,16 , competitive selection among potential splice sites (splice site usage prediction) 11,16 and prediction of specific introns (splice junction…”
Detection of nonneutral substitution rates on mammalian phylogenies
Pollard, K. S., Hubisz, M. J., Rosenbloom, K. R., & Siepel, A. (2010) · Genome Research · doi:10.1101/gr.097857.109 · Cited by 2,353 · cited 1×
“The phyloP 62 score for this position is −3.29, consistent with the fact that gain-of-function variants can occur in fast-evolving parts of the genome.”
InfluentialPredicting effects of noncoding variants with deep learning–based sequence model
Zhou, J., & Troyanskaya, O. G. (2015) · Nature Methods · doi:10.1038/nmeth.3547 · Cited by 2,338
Accurate proteome-wide missense variant effect prediction with AlphaMissense
Cheng, J., Novati, G., Pan, J., Bycroft, C., Žemgulytė, A., Applebaum, T., Pritzel, A., Wong, L. H., Zielinski, M., Sargeant, T., et al. (2023) · Science · doi:10.1126/science.adg7492 · Cited by 2,163
Self-citationRoFormer: Enhanced transformer with rotary position embedding
Su, J., Ahmed, M., Lu, Y., Pan, S., Bo, W., & Liu, Y. (2024) · Neurocomputing · doi:10.1016/j.neucom.2023.127063 · Cited by 1,957
Plasma proteomic associations with genetics and health in the UK Biobank
Sun, B. B., Chiou, J., Traylor, M., Benner, C., Hsu, Y.-H., Richardson, T. G., Surendran, P., Mahajan, A., Robins, C., Vasquez-Grinnell, S. G., et al. (2023) · Nature · doi:10.1038/s41586-023-06592-6 · Cited by 1,880 · cited 1×
“Additionally, the variant is a plasma pQTL for CELSR2, as well as a GTEx eQTL for CELSR2 and PSRC1 17,65 .”
TERT Promoter Mutations in Familial and Sporadic Melanoma
Horn, S., Figl, A., Rachakonda, P. S., Fischer, C., Sucker, A., Gast, A., Kadel, S., Moll, I., Nagore, E., Hemminki, K., et al. (2013) · Science · doi:10.1126/science.1230062 · Cited by 1,799
De novo assembly of bacterial transcriptomes from RNA-seq data
Tjaden, B. C. (2015) · Genome biology · doi:10.1186/s13059-014-0572-2 · Cited by 1,700
Computationally efficient whole-genome regression for quantitative and binary traits
Mbatchou, J., Barnard, L., Backman, J., Marcketta, A., Kosmicki, J. A., Ziyatdinov, A., Benner, C., O’Dushlaine, C., Barber, M., Boutkov, B., et al. (2021) · Nature Genetics · doi:10.1038/s41588-021-00870-7 · Cited by 1,502 · cited 1×
“( h ) AlphaGenome predictions for the variant rs570639864, previously found to be associated with reduced bone mineral density 66 .”
Effective gene expression prediction from sequence by integrating long-range interactions
Avsec, Ž., Agarwal, V., Visentin, D., Ledsam, J. R., Grabska-Barwinska, A., Taylor, K. R., Assael, Y., Jumper, J., Kohli, P., & Kelley, D. R. (2021) · Nature Methods · doi:10.1038/s41592-021-01252-x · Cited by 1,326
Self-citationAlternative polyadenylation of mRNA precursors
Tian, B., & Manley, J. L. (2017) · Nature Reviews Molecular Cell Biology · doi:10.1038/nrm.2016.116 · Cited by 1,179 · cited 1×
“APA is a process that generates transcript diversity by varying the 3′ untranslated region (3′ UTR) of mRNA molecules, often with concomitant impacts on RNA half-life and tissue specificity 28 .”
Splicing regulation: From a parts list of regulatory elements to an integrated splicing code
Wang, Z., & Burge, C. B. (2008) · RNA · doi:10.1261/rna.876308 · Cited by 1,127 · cited 1×
“For example, ISM analysis of exon 9 of the U2SURP gene and its flanking introns high-lighted recognizable splicing-related sequence motifs 18,19 (Fig.”
A simple new approach to variable selection in regression, with application to genetic fine mapping
Wang, G., Sarkar, A., Carbonetto, P., & Stephens, M. (2020) · Journal of the Royal Statistical Society Series B (Statistical Methodology) · doi:10.1111/rssb.12388 · Cited by 1,121
Activity-by-contact model of enhancer–promoter regulation from thousands of CRISPR perturbations
Fulco, C. P., Nasser, J., Jones, T. R., Munson, G., Bergman, D. T., Subramanian, V., Grossman, S. R., Anyoha, R., Doughty, B. R., Patwardhan, T. A., et al. (2019) · Nature Genetics · doi:10.1038/s41588-019-0538-0 · Cited by 1,111
JASPAR 2024: 20th anniversary of the open-access database of transcription factor binding profiles
Rauluševičiūtė, I., Puig, R. R., Blanc‐Mathieu, R., Castro-Mondragon, J. A., Ferenc, K., Kumar, V., Lemma, R. B., Lucas, J., Chèneby, J., Baranas̆ić, D., et al. (2023) · Nucleic Acids Research · doi:10.1093/nar/gkad1059 · Cited by 1,110
Kelley, D. R., Snoek, J., & Rinn, J. L. (2016) · Genome Research · doi:10.1101/gr.200535.115 · Cited by 1,048
The role of regulatory variation in complex traits and disease
Albert, F. W., & Kruglyak, L. (2015) · Nature Reviews Genetics · doi:10.1038/nrg3891 · Cited by 1,047 · cited 1×
“Beyond impacting isoform composition through splicing modulation, non-coding variants can influence traits and cause diseases by altering gene expression 23,24 .”
Predictive Identification of Exonic Splicing Enhancers in Human Genes
Fairbrother, W. G., Yeh, R., Sharp, P. A., & Burge, C. B. (2002) · Science · doi:10.1126/science.1073774 · Cited by 1,020 · cited 1×
“For example, ISM analysis of exon 9 of the U2SURP gene and its flanking introns high-lighted recognizable splicing-related sequence motifs 18,19 (Fig.”
New developments on the Encyclopedia of DNA Elements (ENCODE) data portal
Luo, Y., Hitz, B. C., Gabdank, I., Hilton, J. A., Kagda, M. S., Lam, B. R., Myers, Z., Sud, P., Jou, J., Lin, K. Z., et al. (2019) · Nucleic Acids Research · doi:10.1093/nar/gkz1062 · Cited by 1,007
The power of genetic diversity in genome-wide association studies of lipids
Graham, S. E., Clarke, S. L., Wu, K.-H. H., Kanoni, S., Zajac, G. J. M., Ramdas, S., Surakka, I., Ntalla, I., Vedantam, S., Winkler, T. W., et al. (2021) · Nature · doi:10.1038/s41586-021-04064-3 · Cited by 972 · cited 1×
“( g ) AlphaGenome predictions for the variant rs12740374, previously found to be associated with low-density lipoprotein cholesterol levels 64 .”
The genomic landscape of pediatric and young adult T-lineage acute lymphoblastic leukemia
Liu, Y., Easton, J., Shao, Y., Maciaszek, J., Wang, Z., Wilkinson, M. R., McCastlain, K., Edmonson, M., Pounds, S. B., Shi, L., et al. (2017) · Nature Genetics · doi:10.1038/ng.3909 · Cited by 918 · cited 3×
“For the oncogenic mutation chr. 1: 47239296: C>ACG 6 , AlphaGenome predicted increases in the activating histone marks H3K27ac and H3K4me1 at the variant, consistent with experimentally observed neo-enhancer formation at that position 6,33 (Fig.”
scMC learns biological variation through the alignment of multiple single-cell genomics datasets
Zhang, L., & Nie, Q. H. (2021) · Genome biology · doi:10.1186/s13059-020-02238-2 · Cited by 861
An oncogenic super-enhancer formed through somatic mutation of a noncoding intergenic element
Mansour, M. R., Abraham, B. J., Anders, L., Berezovskaya, A., Gutierrez, A., Durbin, A. D., Etchin, J., Lawton, L., Sallan, S. E., Silverman, L. B., et al. (2014) · Science · doi:10.1126/science.1259037 · Cited by 801
Nucleotide Transformer: building and evaluating robust foundation models for human genomics
Dalla-Torre, H., Gonzalez, L., Mendoza-Revilla, J., Lopez Carranza, N., Grzywaczewski, A. H., Oteri, F., Dallago, C., Trop, E., de Almeida, B. P., Sirelkhatim, H., et al. (2025) · Nature Methods · doi:10.1038/s41592-024-02523-z · Cited by 772 · cited 1×
“…incorporating a broader range of data modalities (such as DNA methylation and RNA structural features) and pursuing foundational model improvements (such as leveraging DNA language models 45–47 , expanding multi-species capabilities and developing robust methods for assay bias correction 29,48 ).”
Uberon, an integrative multi-species anatomy ontology
Mungall, C. J., Torniai, C., Gkoutos, G. V., Lewis, S. E., & Haendel, M. A. (2012) · Genome biology · doi:10.1186/gb-2012-13-1-r5 · Cited by 732
Genome-wide enhancer maps link risk variants to disease genes
Nasser, J., Bergman, D. T., Fulco, C. P., Guckelberger, P., Doughty, B. R., Patwardhan, T. A., Jones, T. R., Nguyen, T. H., Ulirsch, J. C., Lekschas, F., et al. (2021) · Nature · doi:10.1038/s41586-021-03446-x · Cited by 669
Base-resolution models of transcription-factor binding reveal soft motif syntax
Avsec, Ž., Weilert, M., Shrikumar, A., Krueger, S., Alexandari, A., Dalal, K., Fropf, R., McAnany, C., Gagneur, J., Kundaje, A., et al. (2021) · Nature Genetics · doi:10.1038/s41588-021-00782-6 · Cited by 610
Mountjoy, E., Schmidt, E. M., Carmona, M., Schwartzentruber, J., Peat, G., Miranda, A., Fumis, L., Hayhurst, J., Buniello, A., Karim, M. A., et al. (2021) · Nature Genetics · doi:10.1038/s41588-021-00945-5 · Cited by 581
Modeling sample variables with an Experimental Factor Ontology
Malone, J., Holloway, E., Adamusiak, T., Kapushesky, M., Zheng, J., Kolesnikov, N. N., Zhukova, A., Brāzma, A., & Parkinson, H. (2010) · Bioinformatics · doi:10.1093/bioinformatics/btq099 · Cited by 570
Sequential regulatory activity prediction across chromosomes with convolutional neural networks
Kelley, D. R., Reshef, Y. A., Bileschi, M., Belanger, D., McLean, C. Y., & Snoek, J. (2018) · Genome Research · doi:10.1101/gr.227819.117 · Cited by 537
Open Targets: a platform for therapeutic target identification and validation
Koscielny, G., An, P., Carvalho‐Silva, D., Cham, J. A., Fumis, L., Gasparyan, R., Hasan, S., Karamanis, N., Maguire, M., Papa, E., et al. (2016) · Nucleic Acids Research · doi:10.1093/nar/gkw1055 · Cited by 509
Trevino, A. E., Müller, F., Andersen, J., Sundaram, L., Kathiria, A., Shcherbina, A., Farh, K., Chang, H. Y., Pașca, A. M., Kundaje, A., et al. (2021) · Cell · doi:10.1016/j.cell.2021.07.039 · Cited by 478
The sequences of 150,119 genomes in the UK Biobank
Halldorsson, B. V., Eggertsson, H. P., Moore, K. H. S., Hauswedell, H., Eiriksson, O., Ulfarsson, M. O., Palsson, G., Hardarson, M. T., Oddsson, A., Jensson, B. O., et al. (2022) · Nature · doi:10.1038/s41586-022-04965-x · Cited by 448 · cited 1×
“Given that more than 98% of observed genetic variation in humans is non-coding 7 , global characterization of the complex effects of this vast majority of variants remains intractable without computational predictions.”
A compendium of uniformly processed human gene expression and splicing quantitative trait loci
Kerimov, N., Hayhurst, J. D., Peikova, K., Manning, J. R., Walter, P., Kolberg, L., Samoviča, M., Sakthivel, M. P., Kuzmin, I., Trevanion, S. J., et al. (2021) · Nature Genetics · doi:10.1038/s41588-021-00924-w · Cited by 440
Are splicing mutations the most frequent cause of hereditary disease?
López-Bigas, N., Audit, B., Ouzounis, C., Parra, G., & Guigó, R. (2005) · FEBS Letters · doi:10.1016/j.febslet.2005.02.047 · Cited by 412 · cited 1×
“One of the main ways genetic variants cause disease is by disrupting splicing 15 , a process that produces mature RNA sequences by excising”
UniPROBE: an online database of protein binding microarray data on protein-DNA interactions
Newburger, D. E., & Bulyk, M. L. (2008) · Nucleic Acids Research · doi:10.1093/nar/gkn660 · Cited by 376 · cited 1×
“Myb motif from a previous study 6 , originally from UniPROBE 54 . f , Multimodality in trait-altering non-coding variants.”
The Cell Ontology 2016: enhanced content, modularization, and ontology interoperability
Diehl, A. D., Meehan, T. F., Bradford, Y. M., Brush, M., Dahdul, W., Dougall, D. S., He, Y. O., Osumi-Sutherland, D., Ruttenberg, A., Sarntivijai, S., et al. (2016) · Journal of Biomedical Semantics · doi:10.1186/s13326-016-0088-7 · Cited by 332
Predicting 3D genome folding from DNA sequence with Akita
Fudenberg, G., Kelley, D. R., & Pollard, K. S. (2020) · Nature Methods · doi:10.1038/s41592-020-0958-x · Cited by 300
Predicting RNA splicing from DNA sequence using Pangolin
Zeng, T., & Li, Y. I. (2022) · Genome Biology · doi:10.1186/s13059-022-02664-4 · Cited by 283 · cited 2×
“…outcomes can be modelled at three levels: the probability that any given nucleotide acts as a splice donor or acceptor (splice site prediction) 4,11,16 , competitive selection among potential splice sites (splice site usage prediction) 11,16 and prediction of specific introns (splice junction…”
Positional cloning of a novel gene on chromosome 16q causing Bardet-Biedl syndrome (BBS2)
Nishimura, D. Y. (2001) · Human Molecular Genetics · doi:10.1093/hmg/10.8.865 · Cited by 271
A sequence-based global map of regulatory activity for deciphering human genetics
Chen, K. M., Wong, A. K., Troyanskaya, O. G., & Zhou, J. (2022) · Nature Genetics · doi:10.1038/s41588-022-01102-2 · Cited by 269
Predicting RNA-seq coverage from DNA sequence as a unifying model of gene regulation
Linder, J., Srivastava, D., Yuan, H., Agarwal, V., & Kelley, D. R. (2025) · Nature Genetics · doi:10.1038/s41588-024-02053-6 · Cited by 265 · cited 1×
“AlphaGenome performed best on fine-mapped splicing QTL (sQTL) classification 2,20 , including both single-nucleotide polymorphisms (SNPs) within 10 kb of the closest splice site and proximal variants within 200 bp of splice sites (Fig.”
Spectrum of Mutations in Gitelman Syndrome
Vargas‐Poussou, R., Dahan, K., Kahila, D., Vénisse, A., Riveira‐Muñoz, E., Debaix, H., Grisart, B., Bridoux, F., Unwin, R. J., Moulin, B., et al. (2011) · Journal of the American Society of Nephrology · doi:10.1681/asn.2010090907 · Cited by 241
Wang, R., Nambiar, R., Zheng, D., & Tian, B. (2018) · Nucleic Acids Research · doi:10.1093/nar/gkx1000 · Cited by 239
Kircher, M., Xiong, C., Martin, B., Schubach, M., Inoue, F., Bell, R. J. A., Costello, J. F., Shendure, J., & Ahituv, N. (2019) · Nature Communications · doi:10.1038/s41467-019-11526-w · Cited by 234 · cited 1×
“We therefore assessed the performance of AlphaGenome on the 5th Critical Assessment of Genome Interpretation (CAGI5) saturation mutagenesis massively parallel reporter assay (MPRA) challenge 31,32 .”
Genome modeling and design across all domains of life with Evo 2Preprint
Brixi, G., Durrant, M. G., Ku, J., Poli, M., Brockman, G., Chang, D., Gonzalez, G. A., King, S. B., Li, D. B., Merchant, A. T., et al. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.02.18.638918 · Cited by 218 · cited 1×
“Finally, AlphaGenome can complement the capabilities of generative models trained on DNA sequences by predicting functional properties of newly generated sequences 39 .”
Pampari, A., Shcherbina, A., Kvon, E. Z., Kosicki, M., Nair, S., Kundu, S., Kathiria, A. S., Risca, V. I., Kuningas, K., Alasoo, K., et al. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.12.25.630221 · Cited by 183
The 4D Nucleome Data Portal as a resource for searching and visualizing curated nucleomics data
Reiff, S. B., Schroeder, A. J., Kırlı, K., Cosolo, A., Bakker, C., Mercado, L., Lee, S., Veit, A. D., Balashov, A. K., Vitzthum, C. M., et al. (2022) · Nature Communications · doi:10.1038/s41467-022-29697-4 · Cited by 177
Sequence-based modeling of three-dimensional genome architecture from kilobase to chromosome scale
Zhou, J. (2022) · Nature Genetics · doi:10.1038/s41588-022-01065-4 · Cited by 177
Cell-type-directed design of synthetic enhancers
Taskiran, I. I., Spanier, K. I., Dickmänken, H., Kempynck, N., Pančíková, A., Ekşi, E. C., Hulselmans, G., Ismail, J. N., Theunis, K., Vandepoel, R., et al. (2024) · Nature · doi:10.1038/s41586-023-06936-2 · Cited by 175 · cited 1×
“The improved splicing, expression and accessibility predictions of AlphaGenome could be used ‘in the loop’ to accelerate sequence design applications, such as therapeutic antisense oligonucleotides 36 and tissue-specific enhancers 37,38 .”
A harmonized public resource of deeply sequenced diverse human genomesPreprint
Koenig, Z., Yohannes, M. T., Nkambule, L. L., Zhao, X., Goodrich, J. K., Kim, H. A., Wilson, M. W., Tiao, G., Hao, S. P., Sahakian, N., et al. (2024) · Genome Research · doi:10.1101/gr.278378.123 · Cited by 168
Detection of aberrant gene expression events in RNA sequencing data
Yépez, V. A., Mertes, C., Müller, M., Klaproth-Andrade, D., Wachutka, L., Frésard, L., Gušić, M., Scheller, I. F., Goldberg, P. F., Prokisch, H., et al. (2021) · Nature Protocols · doi:10.1038/s41596-020-00462-5 · Cited by 163
DNA language models are powerful predictors of genome-wide variant effects
Benegas, G., Batra, S. S., & Song, Y. S. (2023) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.2311219120 · Cited by 157
Quantifying the regulatory effect size of cis -acting genetic variation using allelic fold changePreprint
Mohammadi, P., Castel, S. E., Brown, A. A., & Lappalainen, T. (2017) · Genome Research · doi:10.1101/gr.216747.116 · Cited by 150
WNT7B Promotes Bone Formation in part through mTORC1
Chen, J., Tu, X., Esen, E., Joeng, K. S., Lin, C., Arbeit, J. M., Rüegg, M. A., Hall, M. N., Ma, L., & Long, F. (2014) · PLoS Genetics · doi:10.1371/journal.pgen.1004145 · Cited by 141
Sasse, A., Ng, B., Spiro, A. E., Tasaki, S., Bennett, D. A., Gaiteri, C., De Jager, P. L., Chikina, M., & Mostafavi, S. (2023) · Nature Genetics · doi:10.1038/s41588-023-01524-6 · Cited by 139 · cited 1×
“We have not yet benchmarked the model on personal genome prediction, which is a known weakness of models in this space 40,41 .”
Chong, R., Insigne, K. D., Yao, D., Burghard, C. P., Wang, J., Hsiao, Y.-H. E., Jones, E. M., Goodman, D. B., Xiao, X., & Kosuri, S. (2019) · Molecular Cell · doi:10.1016/j.molcel.2018.10.037 · Cited by 130 · cited 1×
“…region (within 6 bp intronic or 3 bp exonic) and missense variants predicted as ‘likely_benign’ by AlphaMissense 51 . i , MFASS splicing variant classification (MPRA-tested variants). auPRC on the classification of experimentally validated splice-disrupting variants (data from Chong et al. 22 ).”
Karollus, A., Mauermeier, T., & Gagneur, J. (2023) · Genome Biology · doi:10.1186/s13059-023-02899-9 · Cited by 128
Self-citationSarntivijai, S., Lin, Y., Xiang, Z., Meehan, T. F., Diehl, A. D., Vempati, U. D., Schürer, S. C., Pang, C., MALONE, J. M., Parkinson, H., et al. (2014) · Journal of Biomedical Semantics · doi:10.1186/2041-1480-5-37 · Cited by 127
Personal transcriptome variation is poorly explained by current genomic deep learning models
Huang, C., Shuai, R. W., Baokar, P., Chung, R., Rastogi, R., Kathail, P., & Ioannidis, N. M. (2023) · Nature Genetics · doi:10.1038/s41588-023-01574-w · Cited by 127 · cited 1×
“We have not yet benchmarked the model on personal genome prediction, which is a known weakness of models in this space 40,41 .”
Pooled ChIP-Seq Links Variation in Transcription Factor Binding to Complex Disease Risk
Tehranchi, A. K., Myrthil, M., Martin, T., Hie, B. L., Golan, D., & Fraser, H. B. (2016) · Cell · doi:10.1016/j.cell.2016.03.041 · Cited by 120 · cited 1×
“To illustrate this performance and gain mechanistic insight, we examined specific examples, including fine-mapped caQTLs (African ancestry) and SPI1 bQTLs 30 .”
An encyclopedia of enhancer-gene regulatory interactions in the human genome
Gschwind, A. R., Mualim, K. S., Karbalayghareh, A., Sheth, M. U., Dey, K. K., Jagoda, E., Nurtdinov, R. N., Xi, W., Tan, A. S., Jones, H., et al. (2023) · doi:10.1101/2023.11.09.563812 · Cited by 119 · cited 4×
“ENCODE-rE2G is a logistic regression model trained to predict enhancer-gene interactions from features 12 .”
InfluentialSelf-citationASCOT identifies key regulators of neuronal subtype-specific splicing
Ling, J. P., Wilks, C., Charles, R., Leavey, P. J., Ghosh, D., Jiang, L., Santiago, C. P., Pang, B., Venkataraman, A., Clark, B. S., et al. (2020) · Nature Communications · doi:10.1038/s41467-019-14020-5 · Cited by 107
Batch Normalization Biases Residual Blocks Towards the Identity Function in Deep NetworksPreprint
De, S., & Smith, S. L. (2020) · arXiv (Cornell University) · doi:10.48550/arxiv.2002.10444 · Cited by 97
Self-citationMultiscale footprints reveal the organization of cis-regulatory elements
Yan, H., Horlbeck, M. A., Zhang, R., Ma, S., Shrestha, R., Kartha, V. K., Duarte, F. M., Hock, C., Savage, R. E., Labade, A. S., et al. (2025) · Nature · doi:10.1038/s41586-024-08443-4 · Cited by 93 · cited 1×
“…incorporating a broader range of data modalities (such as DNA methylation and RNA structural features) and pursuing foundational model improvements (such as leveraging DNA language models 45–47 , expanding multi-species capabilities and developing robust methods for assay bias correction 29,48 ).”
Aberrant splicing prediction across human tissues
Wagner, N., Çelik, M. H., Hölzlwimmer, F. R., Mertes, C., Prokisch, H., Yépez, V. A., & Gagneur, J. (2023) · Nature Genetics · doi:10.1038/s41588-023-01373-3 · Cited by 92
Targeted design of synthetic enhancers for selected tissues in the Drosophila embryo
de Almeida, B. P., Schaub, C., Pagani, M., Secchia, S., Furlong, E. E. M., & Stark, A. (2024) · Nature · doi:10.1038/s41586-023-06905-9 · Cited by 87 · cited 1×
“The improved splicing, expression and accessibility predictions of AlphaGenome could be used ‘in the loop’ to accelerate sequence design applications, such as therapeutic antisense oligonucleotides 36 and tissue-specific enhancers 37,38 .”
The spectrum of beta-thalassemia genes in China and Southeast Asia
Kazazian, H. J., Dowling, C., Waber, P. G., Huang, S. S.-L., & Wh, L. (1986) · Blood · doi:10.1182/blood.v68.4.964.964 · Cited by 84
Kaneko, K., Furuyama, K., Fujiwara, T., Kobayashi, R., Ishida, H., Harigae, H., & Shibahara, S. (2013) · Haematologica · doi:10.3324/haematol.2013.085449 · Cited by 83 · cited 1×
“( d ) AlphaGenome predictions for the variant chrX:55028202:A>G, previously reported to result in reduced GATA binding and reduced expression of ALAS2 , thereby possibly causing Sideroblastic Anemia 60 .”
Yuregir, G., Aksoy, K., Çürük, M. A., Dikmen, N., Fei, Y. J., BAYSAL, E., & Huisman, T. H. J. (1992) · British Journal of Haematology · doi:10.1111/j.1365-2141.1992.tb04568.x · Cited by 72
Bolduc, V., Foley, A. R., Solomon-Degefa, H., Sarathy, A., Donkervoort, S. C., Hu, Y., Chen, G. S., Sizov, K., Nalls, M., Zhou, H., et al. (2019) · JCI Insight · doi:10.1172/jci.insight.124403 · Cited by 68
Species-aware DNA language models capture regulatory elements and their evolution
Karollus, A., Hingerl, J., Gankin, D., Grosshauser, M., Klemon, K., & Gagneur, J. (2024) · Genome biology · doi:10.1186/s13059-024-03221-x · Cited by 66
Self-citationGenetic effects on promoter usage are highly context-specific and contribute to complex traits
Alasoo, K., Rodrigues, J., Danesh, J., Freitag, D. F., Paul, D. S., & Gaffney, D. J. (2019) · eLife · doi:10.7554/elife.41673 · Cited by 65
Discovery of regulatory noncoding variants in individual cancer genomes by using cis-X
Liu, Y., Li, C., Shen, S., Chen, X., Szlachta, K., Edmonson, M. N., Shao, Y., Ma, X., Hyle, J., Wright, S., et al. (2020) · Nature Genetics · doi:10.1038/s41588-020-0659-5 · Cited by 64 · cited 2×
“We evaluated three groups of gain-of-function mutations affecting the TAL1 gene in T cell acute lymphoblastic leukaemia (T-ALL): a cluster of 5′ neo-enhancer mutations upstream of the TAL1 TSS 6,33 , an intronic single-nucleotide variant (SNV) 34 and a 3′ neo-enhancer 35 .”
Improved detection of aberrant splicing with FRASER 2.0 and the intron Jaccard index
Scheller, I. F., Lutz, K., Mertes, C., Yépez, V. A., & Gagneur, J. (2023) · The American Journal of Human Genetics · doi:10.1016/j.ajhg.2023.10.014 · Cited by 59
Kerimov, N., Tambets, R., Hayhurst, J. D., Rahu, I., Kolberg, P., Raudvere, U., Kuzmin, I., Chowdhary, A., Vija, A., Teras, H., et al. (2023) · PLoS Genetics · doi:10.1371/journal.pgen.1010932 · Cited by 57
Shigaki, D., Adato, O., Adhikari, A. N., Dong, S., Hawkins‐Hooker, A., Inoue, F., Juven‐Gershon, T., Kenlay, H., Martin, B., Patra, A., et al. (2019) · Human Mutation · doi:10.1002/humu.23797 · Cited by 54
Deciphering the impact of genetic variation on human polyadenylation using APARENT2
Linder, J., Koplik, S. E., Kundaje, A., & Seelig, G. (2022) · Genome Biology · doi:10.1186/s13059-022-02799-4 · Cited by 49
scooby: modeling multimodal genomic profiles from DNA sequence at single-cell resolution
Hingerl, J., Martens, L. D., Karollus, A., Manz, T., Buenrostro, J. D., Theis, F. J., & Gagneur, J. (2025) · Nature Methods · doi:10.1038/s41592-025-02854-5 · Cited by 46
Self-citationMatsunaga, A., Sasaki, J., Han, H., Huang, W., Kugi, M., Koga, T., Ichiki, S., Shinkawa, T., & Arakawa, K. (1999) · Arteriosclerosis Thrombosis and Vascular Biology · doi:10.1161/01.atv.19.2.348 · Cited by 45 · cited 1×
“( a ) AlphaGenome predictions for the variant chr11:116837649:T>G, previously reported to result in underexpression of APOA1 and Hypoalphalipoproteinemia 57 .”
Gillis, E., Kempers, M. J. E., Salemink, S., Timmermans, J., Cheriex, E. C., Bekkers, S. C. A. M., Fransén, E., de Die-Smulders, C. E. M., Loeys, B., & Laer, L. V. (2014) · Human Mutation · doi:10.1002/humu.22540 · Cited by 44
Collins, F. S., Boehm, C., Waber, P. G., Cj, S., Weissman, S., Forget, B. G., & Kazazian, H. J. (1984) · Blood · doi:10.1182/blood.v64.6.1292.1292 · Cited by 38
An RNA foundation model enables discovery of disease mechanisms and candidate therapeuticsPreprint
Celaj, A., Gao, A. J., Lau, T., Holgersen, E. M., Lo, A., Lodaya, V., Cole, C. B., Denroche, R. E., Spickett, C., Wagih, O., et al. (2023) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2023.09.20.558508 · Cited by 36 · cited 1×
“The improved splicing, expression and accessibility predictions of AlphaGenome could be used ‘in the loop’ to accelerate sequence design applications, such as therapeutic antisense oligonucleotides 36 and tissue-specific enhancers 37,38 .”
Dissecting the cis-regulatory syntax of transcription initiation with deep learningPreprint
Cochran, K., Yin, M., Mantripragada, A., Schreiber, J. M., Marinov, G. K., Shah, S. R., Yu, H., Lis, J. T., & Kundaje, A. B. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.05.28.596138 · Cited by 36
Quantifying splice-site usage: a simple yet powerful approach to analyze splicing
Dent, C., Singh, S., Mukherjee, S., Mishra, S., Sarwade, R. D., Shamaya, N., Loo, K. P., Harrison, P. F., Sureshkumar, S., Powell, D., et al. (2021) · NAR Genomics and Bioinformatics · doi:10.1093/nargab/lqab041 · Cited by 34
Splam: a deep-learning-based splice site predictor that improves spliced alignments
Chao, K.-H., Mao, A., Salzberg, S. L., & Pertea, M. (2024) · Genome biology · doi:10.1186/s13059-024-03379-4 · Cited by 34
Decoding sequence determinants of gene expression in diverse cellular and disease states
Lal, A., Karollus, A., Gunsalus, L. M., Garfield, D., Nair, S., Tseng, A. M., Gordon, M. G., Blischak, J., Geijn, B. V. D., Bhangale, T., et al. (2026) · Nature Methods · doi:10.1038/s41592-026-03102-0 · Cited by 34 · cited 1×
“…accuracy and utility (such as through task-specific calibration, fine-tuning on perturbational datasets or integration of single-cell data 43,44 ), incorporating a broader range of data modalities (such as DNA methylation and RNA structural features) and pursuing foundational model…”
Self-citationHeibel, S. K., Mew, N. A., Caldovic, L., Daikhin, Y., Yudkoff, M., & Tuchman, M. (2011) · Human Mutation · doi:10.1002/humu.21553 · Cited by 32 · cited 1×
“( c ) AlphaGenome predictions for the variant chr17:44001600:C>A, previously reported to result in reduced binding of HNF1, underexpression of NAGS and ultimately N-acetylglutamate Synthase Deficiency 59 .”
TAL1 activation in T-cell acute lymphoblastic leukemia: a novel oncogenic 3’ neo-enhancer
Smith, C., Goyal, A., Weichenhan, D., Allemand, E., Mayakonda, A., Toprak, U., Riedel, A., Balducci, E., Manojkumar, M., Pejkovska, A., et al. (2023) · Haematologica · doi:10.3324/haematol.2022.281583 · Cited by 30 · cited 2×
“We evaluated three groups of gain-of-function mutations affecting the TAL1 gene in T cell acute lymphoblastic leukaemia (T-ALL): a cluster of 5′ neo-enhancer mutations upstream of the TAL1 TSS 6,33 , an intronic single-nucleotide variant (SNV) 34 and a 3′ neo-enhancer 35 .”
Benchmarking DNA Sequence Models for Causal Regulatory Variant Prediction in Human GeneticsPreprint
Benegas, G., Eraslan, G., & Song, Y. S. (2025) · bioRxiv · doi:10.1101/2025.02.11.637758 · Cited by 29 · cited 1×
“Fraction of trait-affecting variants 55 which is disrupted by the variant, thereby leading to an aberrant transcript and reduced expression (Fig.”
ATP7B variant c.1934T > G p.Met645Arg causes Wilson disease by promoting exon 6 skipping
Merico, D., Spickett, C., O’Hara, M., Kakaradov, B., Deshwar, A. G., Fradkin, P., Gandhi, S., Gao, J., Grant, S., Kron, K. J., et al. (2020) · npj Genomic Medicine · doi:10.1038/s41525-020-0123-6 · Cited by 25
Decoding sequence determinants of gene expression in diverse cellular and disease states
Lal, A., Karollus, A., Gunsalus, L., et al. (2024) · doi:10.1101/2024.10.09.617507 · Cited by 23
Self-citationUncertainty-aware genomic deep learning with knowledge distillation
Zhou, J., Rizzo, K., Christensen, T. A., Tang, Z., & Koo, P. K. (2026) · npj Artificial Intelligence · doi:10.1038/s44387-025-00053-3 · Cited by 22
Reference-informed prediction of alternative splicing and splicing-altering mutations from sequencesPreprint
Xu, C., Bao, S., Wang, Y., Li, W., Chen, H., Shen, Y., Jiang, T., & Zhang, C. (2024) · Genome Research · doi:10.1101/gr.279044.124 · Cited by 17 · cited 2×
“…outcomes can be modelled at three levels: the probability that any given nucleotide acts as a splice donor or acceptor (splice site prediction) 4,11,16 , competitive selection among potential splice sites (splice site usage prediction) 11,16 and prediction of specific introns (splice junction…”
Uncertainty-aware genomic deep learning with knowledge distillation
Zhou, J., Rizzo, K., Christensen, T., Tang, Z., & Koo, P. K. (2024) · doi:10.1101/2024.11.13.623485 · Cited by 14
Alternative Splicing and Disease
Jeanteur, P. (2006) · Progress in molecular and subcellular biology · doi:10.1007/978-3-540-34449-0 · Cited by 9
Parameter-Efficient Fine-Tuning of a Supervised Regulatory Sequence ModelPreprint
Yuan, H., Linder, J., & Kelley, D. R. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.05.26.656171 · Cited by 8
Finucane, H. K., Parsa, S., Guez, J., Kanai, M., Satterstrom, F. K., Nkambule, L. L., Daly, M. J., Seed, C., & Karczewski, K. J. (2024) · doi:10.1101/2024.09.17.613327 · Cited by 4 · cited 1×
“…is limited because AlphaGenome predicts molecular consequences of variants, whereas these phenotypes involve broader biological processes (including gene function, development and environmental factors) and gene-to-disease effects 42 beyond the direct sequence-to-function scope of the model.”
Cited by
Sorted by citations
Benchmarking DNA foundation models for genomic and genetic tasks
Feng, H., Wu, L., Zhao, B., Huff, C. D., Zhang, J. J., Wu, J., Lin, L., Wei, P., & Wu, C. (2025) · Nature Communications · doi:10.1038/s41467-025-65823-8 · Cited by 83
Harnessing artificial intelligence to advance CRISPR-based genome editing technologies
Thomson, T., Li, G. L., Strilchuk, A. W., Cui, H., Wang, B., & Li, B. (2025) · Nature Reviews Genetics · doi:10.1038/s41576-025-00907-1 · Cited by 34
Advancing bioinformatics with language models: components, applications, and perspectives
Liu, J., Yang, M., Yu, Y., Xu, H., Wang, T., Li, K., & Zhou, X. (2026) · Briefings in Bioinformatics · doi:10.1093/bib/bbag367 · Cited by 66
Fine-tuning sequence-to-expression models on personal genome and transcriptome data
Rastogi, R., Reddy, A. J., Chung, R., & Ioannidis, N. M. (2026) · Genome biology · doi:10.1186/s13059-026-04091-1 · Cited by 22
CREsted: modeling genomic and synthetic cell-type-specific enhancers across tissues and species
Kempynck, N., Winter, S. D., Blaauw, C. H., Konstantakos, V., Ekşi, E. C., Dieltiens, S., Abaffyová, D., Bercier, V., Taskiran, I. I., Hulselmans, G. J., et al. (2026) · Nature Methods · doi:10.1038/s41592-026-03057-2 · Cited by 29
Metadata
- Authors
- Avsec, Ž., Latysheva, N., Cheng, J., Novati, G., Taylor, K. R., Ward, T., Bycroft, C., Nicolaisen, L., Arvaniti, E., Pan, J., Thomas, R., Dutordoir, V., Perino, M., De, S., Karollus, A., Gayoso, A., Sargeant, T., Mottram, A., Wong, L. H., Drotár, P., Kosiorek, A., Senior, A., Tanburn, R., Applebaum, T., Basu, S., Hassabis, D., & Kohli, P.
- Affiliations
- Google DeepMind (United Kingdom) (GB)
- Corresponding author
- Kohli, Pushmeet
- ORCID
- Avsec, Žiga
- Cheng, Jun
- Novati, Guido
- Taylor, Kyle R.
- Ward, Tom
- Bycroft, Clare
- Nicolaisen, Lauren
- Arvaniti, Eirini
- Thomas, Raina
- Dutordoir, Vincent
- Perino, Matteo
- De, Soham
- Karollus, Alexander
- Gayoso, Adam
- Mottram, Anne
- Wong, Lai Hong
- Kosiorek, Adam
- Senior, Andrew
- Applebaum, Taylor
- Basu, Souradeep
- Hassabis, Demis
- Kohli, Pushmeet
- Venue
- Nature
- Published
- 2026-01-28
- Volume
- 649
- Issue
- 8099
- Pages
- 1206-1218
- ISSN
- 0028-0836 (ISSN-L)
- 1476-4687
- PMC
- PMC12851941
- OpenAlex
- W7125967877
- Semantic Scholar
- c8d5014d457c4d2c6d8acb5558e3d618c46ba735
- Cited by
- 373
- References
- 110
- Citations by year
- 2026: 330
- FWCI
- 598.07
- Type
- journal article
- Language
- en
- Open access
- hybrid
- doi.org
- Locations
- Nature (publishedVersion, cc-by)
- bioRxiv (Cold Spring Harbor Laboratory) (acceptedVersion, cc-by-nc)
- PubMed (publishedVersion)
- Europe PMC (PubMed Central) (submittedVersion, cc-by)
- PubMed Central (submittedVersion, cc-by)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Chromatin Dynamics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genetic Associations and Epidemiology
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genomics and Rare Diseases
- Keywords
- AlphaGenome, variant effect prediction, gene expression prediction, transcription initiation, chromatin accessibility, histone modifications, transcription factor binding, chromatin contact maps, single base pair resolution, regulatory variants, human and mouse genomes
- MeSH terms
- Deep Learning, Animals, Chromatin, Gene Expression Regulation, Humans, Models, Genetic, Proto-Oncogene Proteins, Transcription Factors, Genetic Variation, Genome, Human, Genome, Sequence Analysis, DNA, RNA Splice Sites, Genomics, Mice
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
A language model drew the claims from the paper on 2026-10-20, under version 3 of the claim definition (model deepseek-v4.1-flash, prompt extract-v3).
A second model labelled each claim on 2026-10-20 (model jev-1.13-20260917, definition revision gold-guideline-v0.2).
Cited sources are checked with a language model’s help (gpt-6-sol).
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
