

The DOI matches its Crossref record; 30 of the 31 references it cites were found in a registry or an index.
A coherent approach for analysis of the Illumina HumanMethylation450 BeadChip improves data quality and performance in epigenome-wide association studies
Open accessAbstract
DNA methylation plays a fundamental role in the regulation of the genome, but the optimal strategy for analysis of genome-wide DNA methylation data remains to be determined. We developed a comprehensive analysis pipeline for epigenome-wide association studies (EWAS) using the Illumina Infinium HumanMethylation450 BeadChip, based on 2,687 individuals, with 36 samples measured in duplicate. We propose new approaches to quality control, data normalisation and batch correction through control-probe adjustment and establish a null hypothesis for EWAS using permutation testing. Our analysis pipeline outperforms existing approaches, enabling accurate identification of methylation quantitative trait loci for hypothesis driven follow-up experiments.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
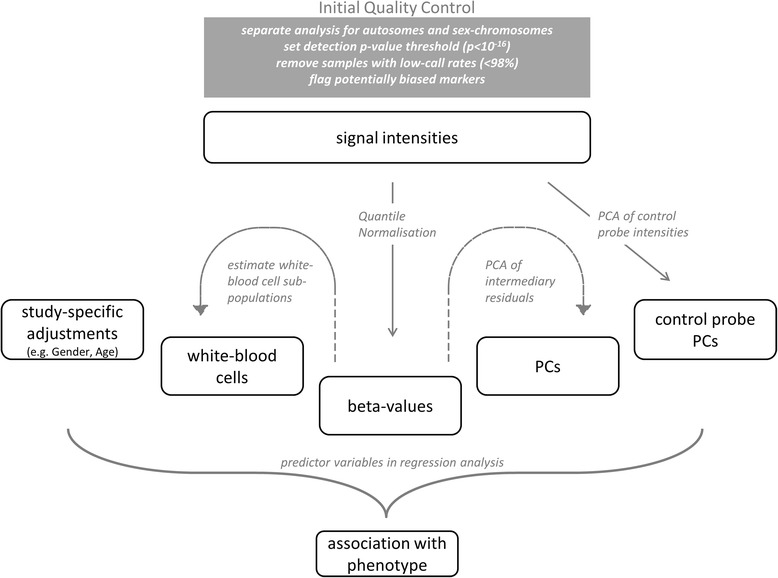
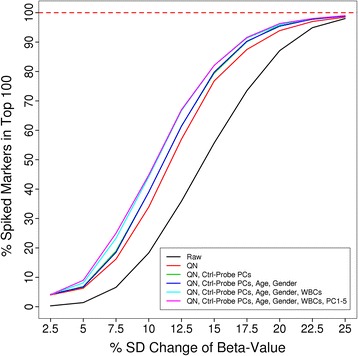
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
In the citing paper
“One of the suggestive CpGs (cg04099436) was flagged for cross-reactivity (cf. Lehne et al., 2015; Pidsley et al., 2016) and thus omitted from the results.”
From Epigenome-wide DNA methylation in unipolar depression: predictive biomarker of antidepressant treatment response? (2024) · cites this paper as Lehne et al., 2015
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Adjusting batch effects in microarray expression data using empirical Bayes methods
Johnson, W. E., Li, C., & Rabinovic, A. (2007) · Biostatistics · doi:10.1093/biostatistics/kxj037 · Cited by 8,658
limma: Linear Models for Microarray Data
Smyth, G. K. (2005) · Statistics for Biology and Health · doi:10.1007/0-387-29362-0_23 · Cited by 5,796
DNA methylation arrays as surrogate measures of cell mixture distribution
Houseman, E. A., Accomando, W. P., Koestler, D. C., Christensen, B. C., Marsit, C. J., Nelson, H. H., Wiencke, J. K., & Kelsey, K. T. (2012) · BMC Bioinformatics · doi:10.1186/1471-2105-13-86 · Cited by 3,175
lumi : a pipeline for processing Illumina microarray
Du, P., Kibbe, W. A., & Lin, S. M. (2008) · Bioinformatics · doi:10.1093/bioinformatics/btn224 · Cited by 2,028
High density DNA methylation array with single CpG site resolution
Bibikova, M., Barnes, B., Tsan, C., Ho, V., Klotzle, B., Le, J. M., Delano, D., Zhang, L., Schroth, G. P., Gunderson, K. L., et al. (2011) · Genomics · doi:10.1016/j.ygeno.2011.07.007 · Cited by 1,727 · cited 6×
“We developed a comprehensive analysis pipeline for epigenome-wide association studies (EWAS) using the Illumina Infinium HumanMethylation450 BeadChip, based on 2,687 individuals, with 36 samples measured in duplicate.”
BackgroundInfluentialTeschendorff, A. E., Marabita, F., Lechner, M., Bartlett, T. E., Tegner, J. N., Gómez-Cabrero, D., & Beck, S. (2012) · Bioinformatics · doi:10.1093/bioinformatics/bts680 · Cited by 1,686
Kulis, M., & Esteller, M. (2010) · Advances in genetics · doi:10.1016/b978-0-12-380866-0.60002-2 · Cited by 1,617
Chen, Y. A., Lemire, M., Choufani, S., Butcher, D. T., Grafodatskaya, D., Zanke, B. W., Gallinger, S., Hudson, T. J., & Weksberg, R. (2013) · Epigenetics · doi:10.4161/epi.23470 · Cited by 1,547
DNA methylation profiling of human chromosomes 6, 20 and 22
Eckhardt, F., Lewin, J., Cortese, R., Rakyan, V. K., Attwood, J., Burger, M., Burton, J. H., Cox, T. V., Davies, R., Down, T. A., et al. (2006) · Nature Genetics · doi:10.1038/ng1909 · Cited by 1,378
A data-driven approach to preprocessing Illumina 450K methylation array data
Pidsley, R., Y Wong, C. C., Volta, M., Lunnon, K., Mill, J., & Schalkwyk, L. C. (2013) · BMC Genomics · doi:10.1186/1471-2164-14-293 · Cited by 1,143 · cited 1×
“On the basis of these results, which are in agreement with previous findings [23,24], we performed quantile normalisation of intensity values for all samples in this study.”
ResultChAMP: 450k Chip Analysis Methylation Pipeline
Morris, T. J., Butcher, L. M., Feber, A., Teschendorff, A. E., Chakravarthy, A., Wojdacz, T. K., & Beck, S. (2013) · Bioinformatics · doi:10.1093/bioinformatics/btt684 · Cited by 1,025
Epigenetic events in mammalian germ-cell development: reprogramming and beyond
Sasaki, H., & Matsui, Y. (2008) · Nature Reviews Genetics · doi:10.1038/nrg2295 · Cited by 904
Functional normalization of 450k methylation array data improves replication in large cancer studies
Fortin, J.-P., Labbe, A., Lemire, M., Zanke, B. W., Hudson, T. J., Fertig, E. J., Greenwood, C. M. T., & Hansen, K. D. (2014) · Genome biology · doi:10.1186/s13059-014-0503-2 · Cited by 893
DNA methylation patterns associate with genetic and gene expression variation in HapMap cell lines
Bell, J. T., Pai, A. A., Pickrell, J. K., Gaffney, D. J., Pique-Regi, R., Degner, J. F., Gilad, Y., & Pritchard, J. K. (2011) · Genome Biology · doi:10.1186/gb-2011-12-1-r10 · Cited by 838
SWAN: Subset-quantile Within Array Normalization for Illumina Infinium HumanMethylation450 BeadChips
Maksimovic, J., Gordon, L., & Oshlack, A. (2012) · Genome Biology · doi:10.1186/gb-2012-13-6-r44 · Cited by 826
Bump hunting to identify differentially methylated regions in epigenetic epidemiology studies
Jaffe, A. E., Murakami, P. N., Lee, H.-J., Leek, J. T., Fallin, M. D., Feinberg, A. P., & Irizarry, R. A. (2012) · International Journal of Epidemiology · doi:10.1093/ije/dyr238 · Cited by 700
Evaluation of the Infinium Methylation 450K technology
Dedeurwaerder, S., Defrance, M., Calonne, E., Denis, H., Sotiriou, C., & Fuks, F. (2011) · Epigenomics · doi:10.2217/epi.11.105 · Cited by 569
Touleimat, N., & Tost, J. (2012) · Epigenomics · doi:10.2217/epi.12.21 · Cited by 517
Price, E. M., Cotton, A. M., Lam, L. L. C., Farré, P., Emberly, E. G., Brown, C. J., Robinson, W. P., & Kobor, M. S. (2013) · Epigenetics & Chromatin · doi:10.1186/1756-8935-6-4 · Cited by 494
Reference-free cell mixture adjustments in analysis of DNA methylation data
Houseman, E. A., Molitor, J., & Marsit, C. J. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu029 · Cited by 441
Volkmar, M., Dedeurwaerder, S., Cunha, D. A., Ndlovu, M. N., Defrance, M., Deplus, R., Calonne, E., Volkmar, U., Igoillo‐Esteve, M., Naamane, N., et al. (2012) · The EMBO Journal · doi:10.1038/emboj.2011.503 · Cited by 420 · cited 1×
“Altered DNA methylation has been linked to complex human diseases including cancer [4], schizophrenia [5], multiple sclerosis [6] and type 2 diabetes [7-9].”
BackgroundToperoff, G., Aran, D., Kark, J. D., Rosenberg, M. H., Dubnikov, T., Nissan, B., Wainstein, J., Friedlander, Y., Levy‐Lahad, E., Gläser, B., et al. (2011) · Human Molecular Genetics · doi:10.1093/hmg/ddr472 · Cited by 353
IMA: an R package for high-throughput analysis of Illumina's 450K Infinium methylation data
Wang, D., Yan, L., Hu, Q., Sucheston, L. E., Higgins, M. J., Ambrosone, C. B., Johnson, C. S., Smiraglia, D. J., & Liu, S. (2012) · Bioinformatics · doi:10.1093/bioinformatics/bts013 · Cited by 309
Epigenome-wide association studies without the need for cell-type composition
Zou, J., Lippert, C., Heckerman, D. E., Aryee, M. J., & Listgarten, J. (2014) · Nature Methods · doi:10.1038/nmeth.2815 · Cited by 204
Dayeh, T. A., Olsson, A. H., Volkov, P., Almgren, P., Rönn, T., & Ling, C. (2013) · Diabetologia · doi:10.1007/s00125-012-2815-7 · Cited by 199
Xist and the order of silencing
Ng, K., Pullirsch, D., Leeb, M., & Wutz, A. (2007) · EMBO Reports · doi:10.1038/sj.embor.7400871 · Cited by 159
Epigenetic changes in patients with multiple sclerosis
Koch, M. W., Metz, L. M., & Kovalchuk, O. (2012) · Nature Reviews Neurology · doi:10.1038/nrneurol.2012.226 · Cited by 142
DNA methylation analysis of BDNF gene promoters in peripheral blood cells of schizophrenia patients
Ikegame, T., Bundo, M., Sunaga, F., Asai, T., Nishimura, F., Yoshikawa, A., Kawamura, Y., Hibino, H., Tochigi, M., Kakiuchi, C., et al. (2013) · Neuroscience Research · doi:10.1016/j.neures.2013.08.004 · Cited by 129
Methylomics of gene expression in human monocytes
Liu, Y., Ding, J., Reynolds, L. M., Lohman, K., Register, T. C., de la Fuente, A., Howard, T. D., Hawkins, G., Cui, W., Morris, J., et al. (2013) · Human Molecular Genetics · doi:10.1093/hmg/ddt356 · Cited by 96
A systematic assessment of normalization approaches for the Infinium 450K methylation platform
Wu, M. C., Joubert, B. R., Kuan, P. F., Håberg, S. E., Nystad, W., Peddada, S. D., & London, S. J. (2013) · Epigenetics · doi:10.4161/epi.27119 · Cited by 78
Cited by
Sorted by citations
DunedinPACE, a DNA methylation biomarker of the pace of aging
Belsky, D. W., Caspi, A., Corcoran, D. L., Sugden, K., Poulton, R., Arseneault, L., Baccarelli, A., Chamarti, K., Gao, X., Hannon, E., et al. (2022) · eLife · doi:10.7554/elife.73420 · Cited by 1,083
Epigenome-wide association study of body mass index, and the adverse outcomes of adiposity
Wahl, S., Drong, A., Lehne, B., Loh, M., Scott, W. R., Kunze, S., Tsai, P.-C., Ried, J. S., Zhang, W., Yang, Y., et al. (2017) · Nature · doi:10.1038/nature20784 · Cited by 945
Self-citationSeSAMe: reducing artifactual detection of DNA methylation by Infinium BeadChips in genomic deletions
Zhou, W., Triche, T. J., Laird, P. W., & Shen, H. (2018) · Nucleic Acids Research · doi:10.1093/nar/gky691 · Cited by 536
Chambers, J. C., Loh, M. C. S., Lehne, B. C., Drong, A., Kriebel, J., Motta, V., Wahl, S., Elliott, H. R., Rota, F., Scott, W. R., et al. (2015) · The Lancet Diabetes & Endocrinology · doi:10.1016/s2213-8587(15)00127-8 · Cited by 526
Self-citationHiggins-Chen, A. T., Thrush, K. L., Wang, Y., Minteer, C. J., Kuo, P.-L., Wang, M., Niimi, P., Sturm, G., Lin, J., Moore, A. Z., et al. (2022) · Nature Aging · doi:10.1038/s43587-022-00248-2 · Cited by 502
Metadata
- Authors
- Lehne, B., Drong, A. W., Loh, M., Zhang, W., Scott, W. R., Tan, S.-T., Afzal, U., Scott, J., Jarvelin, M.-R., Elliott, P., McCarthy, M. I., Kooner, J. S., & Chambers, J. C.
- Affiliations
- Imperial College London (GB)
- Centre for Human Genetics (GB)
- Ealing Hospital NHS Trust (GB)
- MRC Centre for Environment and Health (GB)
- Imperial College Healthcare NHS Trust (GB)
- Corresponding author
- Lehne, Benjamin
- Chambers, John C
- ORCID
- Venue
- Genome biology
- Published
- 2015-02-15
- Volume
- 16
- Issue
- 1
- Pages
- 37-37
- ISSN
- 1465-6906 (ISSN-L)
- 1474-760X
- PMC
- PMC4365767
- OpenAlex
- W4297674379
- Cited by
- 380
- References
- 31
- Citations by year
- 2026: 22 · 2025: 40 · 2024: 42 · 2023: 43 · 2022: 43 · 2021: 54 · 2020: 43 · 2019: 36 · 2018: 26 · 2017: 16 · 2016: 12 · 2015: 2
- FWCI
- 8.11
- Type
- journal article
- Language
- en
- Update notices
- correction (2016-04-21)
- Open access
- gold
- doi.org
- Locations
- Genome biology (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Research Portal (King's College London) (submittedVersion, cc-by)
- Spiral (Imperial College London) (submittedVersion, other-oa)
- Spiral (Imperial College London) (submittedVersion, cc-by-nc-nd)
- STM:n Hallinnonalan avoin julkaisuarkisto (Julkari) (submittedVersion)
- Funders
- Wellcome Trust (10371, 084723/Z/08/Z, 090532, 090532/Z/09/Z, 090532/Z/09/)
- National Institute for Health Research Health Protection Research Unit
- National Institute for Health and Care Research (NF-SI-0611-10136, RP-PG-0407-10371)
- British Heart Foundation (090532/Z/09/Z, SP/04/002)
- Imperial College London
- Imperial College Healthcare NHS Trust
- European Commission (279143)
- MRC-PHE Centre for Environment and Health
- Medical Research Council (G0900747 91070, Hub grant G0900747 91070, MR/L01341X/1, MR/K002414/1, G0601966, G0700931)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Epigenetics and DNA Methylation
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Genetics › Genetic Associations and Epidemiology
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene expression and cancer classification
- Keywords
- DNA methylation, epigenome-wide association study, data quality control, data normalization, batch effect correction, permutation tests, null hypothesis, methylation quantitative trait loci, data analysis pipeline
- MeSH terms
- Humans, Software, Genome, Human, CpG Islands, DNA Methylation, Oligonucleotide Array Sequence Analysis, Quantitative Trait Loci, Epigenesis, Genetic, Genome-Wide Association Study, Epigenomics
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
