

The DOI matches its Crossref record; 76 of the 76 references it cites were found in a registry or an index.
Gaining comprehensive biological insight into the transcriptome by performing a broad-spectrum RNA-seq analysis
Open accessAbstract
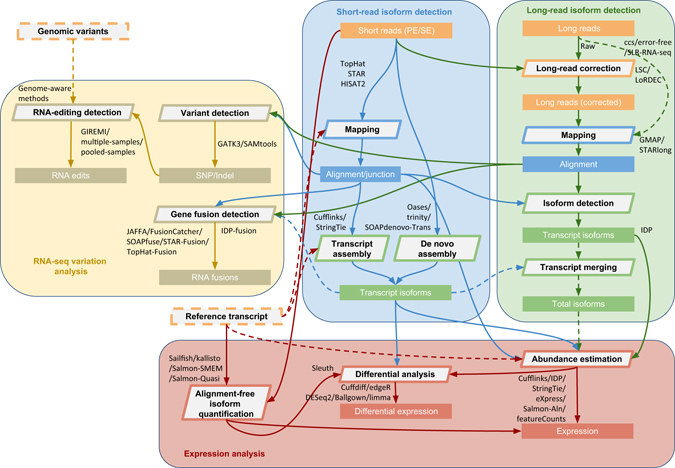
RNA-sequencing (RNA-seq) is an essential technique for transcriptome studies, hundreds of analysis tools have been developed since it was debuted. Although recent efforts have attempted to assess the latest available tools, they have not evaluated the analysis workflows comprehensively to unleash the power within RNA-seq. Here we conduct an extensive study analysing a broad spectrum of RNA-seq workflows. Surpassing the expression analysis scope, our work also includes assessment of RNA variant-calling, RNA editing and RNA fusion detection techniques. Specifically, we examine both short- and long-read RNA-seq technologies, 39 analysis tools resulting in ~120 combinations, and ~490 analyses involving 15 samples with a variety of germline, cancer and stem cell data sets. We report the performance and propose a comprehensive RNA-seq analysis protocol, named RNACocktail, along with a computational pipeline achieving high accuracy. Validation on different samples reveals that our proposed protocol could help researchers extract more biologically relevant predictions by broad analysis of the transcriptome.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“This has been noted by others in the past, and GATK recommendations for variant calling on bulk RNAseq involve removing these regions of the alignments prior to variant calling23.”
From Souporcell: robust clustering of single-cell RNA-seq data by genotype without reference genotypes (2020) · cites this paper as [23]
Experimental — accuracy not yet measured
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Moderated estimation of fold change and dispersion for RNA-seq data with DESeq2
Love, M. I., Huber, W., & Anders, S. (2014) · Genome Biology · doi:10.1186/s13059-014-0550-8 · Cited by 95,106
Trimmomatic: a flexible trimmer for Illumina sequence data
Bolger, A. M., Lohse, M., & Usadel, B. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu170 · Cited by 65,548
The Sequence Alignment/Map format and SAMtools
Li, H., Handsaker, B., Wysoker, A., Fennell, T., Ruan, J., Homer, N., Marth, G., Abecasis, G., Durbin, R., & 1000 Genome Project Data Processing Subgroup (2009) · Bioinformatics · doi:10.1093/bioinformatics/btp352 · Cited by 62,041
STAR: ultrafast universal RNA-seq aligner
Dobin, A., Davis, C. A., Schlesinger, F., Drenkow, J., Zaleski, C., Jha, S., Batut, P., Chaisson, M., & Gingeras, T. R. (2013) · Bioinformatics · doi:10.1093/bioinformatics/bts635 · Cited by 51,709
Robinson, M. D., McCarthy, D. J., & Smyth, G. K. (2010) · Bioinformatics · doi:10.1093/bioinformatics/btp616 · Cited by 41,364
limma powers differential expression analyses for RNA-sequencing and microarray studies
Ritchie, M. E., Phipson, B., Wu, D., Hu, Y., Law, C. W., Shi, W., & Smyth, G. K. (2015) · Nucleic Acids Research · doi:10.1093/nar/gkv007 · Cited by 40,458
featureCounts: an efficient general purpose program for assigning sequence reads to genomic features
Liao, Y., Smyth, G. K., & Shi, W. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btt656 · Cited by 27,219
HISAT: a fast spliced aligner with low memory requirements
Kim, D., Langmead, B., & Salzberg, S. L. (2015) · Nature Methods · doi:10.1038/nmeth.3317 · Cited by 24,112
RSEM: accurate transcript quantification from RNA-Seq data with or without a reference genome
Li, B., & Dewey, C. N. (2011) · BMC Bioinformatics · doi:10.1186/1471-2105-12-323 · Cited by 22,179
Full-length transcriptome assembly from RNA-Seq data without a reference genome
Grabherr, M. G., Haas, B. J., Yassour, M., Levin, J. Z., Thompson, D. A., Amit, I., Adiconis, X., Fan, L., Raychowdhury, R., Zeng, Q., et al. (2011) · Nature Biotechnology · doi:10.1038/nbt.1883 · Cited by 21,201
Trapnell, C., Williams, B. A., Pertea, G., Mortazavi, A., Kwan, G., van Baren, M. J., Salzberg, S. L., Wold, B. J., & Pachter, L. (2010) · Nature Biotechnology · doi:10.1038/nbt.1621 · Cited by 15,556
StringTie enables improved reconstruction of a transcriptome from RNA-seq reads
Pertea, M., Pertea, G. M., Antonescu, C. M., Chang, T.-C., Mendell, J. T., & Salzberg, S. L. (2015) · Nature Biotechnology · doi:10.1038/nbt.3122 · Cited by 14,406
Salmon provides fast and bias-aware quantification of transcript expression
Patro, R., Duggal, G., Love, M. I., Irizarry, R. A., & Kingsford, C. (2017) · Nature Methods · doi:10.1038/nmeth.4197 · Cited by 12,700
Kim, D., Pertea, G., Trapnell, C., Pimentel, H., Kelley, R., & Salzberg, S. L. (2013) · Genome Biology · doi:10.1186/gb-2013-14-4-r36 · Cited by 12,668
Trapnell, C., Roberts, A., Goff, L. A., Pertea, G. M., Kim, D., Kelley, D. R., Pimentel, H., Salzberg, S. L., Rinn, J. L., & Pachter, L. S. (2012) · Nature Protocols · doi:10.1038/nprot.2012.016 · Cited by 12,145
A framework for variation discovery and genotyping using next-generation DNA sequencing data
DePristo, M. A., Banks, E., Poplin, R. E., Garimella, K. V., Maguire, J., Hartl, C., Philippakis, A., del Angel, G., Rivas, M. A., Hanna, M., et al. (2011) · Nature Genetics · doi:10.1038/ng.806 · Cited by 11,508
Near-optimal probabilistic RNA-seq quantification
Bray, N. L., Pimentel, H., Melsted, P., & Pachter, L. (2016) · Nature Biotechnology · doi:10.1038/nbt.3519 · Cited by 9,723
Velvet: Algorithms for de novo short read assembly using de Bruijn graphsPreprint
Zerbino, D. R., & Birney, E. (2008) · Genome Research · doi:10.1101/gr.074492.107 · Cited by 9,587
dbSNP: the NCBI database of genetic variation
Sherry, S. T. (2001) · Nucleic Acids Research · doi:10.1093/nar/29.1.308 · Cited by 7,502
GENCODE: The reference human genome annotation for The ENCODE Project
Harrow, J., Frankish, A., Gonzalez, J. M., Tapanari, E., Diekhans, M., Kokocinski, F., Aken, B. L., Barrell, D., Zadissa, A., Searle, S., et al. (2012) · Genome Research · doi:10.1101/gr.135350.111 · Cited by 4,585
Self-citationDifferential analysis of gene regulation at transcript resolution with RNA-seq
Trapnell, C., Hendrickson, D. G., Sauvageau, M., Goff, L., Rinn, J. L., & Pachter, L. (2013) · Nature Biotechnology · doi:10.1038/nbt.2450 · Cited by 3,492
ToppGene Suite for gene list enrichment analysis and candidate gene prioritization
Chen, J., Bardes, E. E., Aronow, B. J., & Jegga, A. G. (2009) · Nucleic Acids Research · doi:10.1093/nar/gkp427 · Cited by 2,952
A survey of best practices for RNA-seq data analysis
Conesa, A., Madrigal, P., Tarazona, S., Gomez-Cabrero, D., Cervera, A., McPherson, A., Szcześniak, M. W., Gaffney, D. J., Elo, L. L., Zhang, X., et al. (2016) · Genome Biology · doi:10.1186/s13059-016-0881-8 · Cited by 2,794
GMAP: A genomic mapping and alignment program for mRNA and EST sequences
Wu, T. D., & Watanabe, C. K. (2005) · Bioinformatics · doi:10.1093/bioinformatics/bti310 · Cited by 2,354
The Galaxy platform for accessible, reproducible and collaborative biomedical analyses: 2016 update
Afgan, E., Baker, D., van den Beek, M., Blankenberg, D. J., Bouvier, D., Čech, M., Chilton, J. M., Clements, D., Coraor, N., Eberhard, C., et al. (2016) · Nucleic Acids Research · doi:10.1093/nar/gkw343 · Cited by 2,150
Differential analysis of RNA-seq incorporating quantification uncertainty
Pimentel, H., Bray, N. L., Puente, S., Melsted, P., & Pachter, L. (2017) · Nature Methods · doi:10.1038/nmeth.4324 · Cited by 1,534
Oases:robustde novoRNA-seq assembly across the dynamic range of expression levels
Schulz, M. H., Zerbino, D. R., Vingron, M., & Birney, E. (2012) · Bioinformatics · doi:10.1093/bioinformatics/bts094 · Cited by 1,470
dbEST — database for “expressed sequence tags”
Boguski, M. S., Lowe, T. M., & Tolstoshev, C. M. (1993) · Nature Genetics · doi:10.1038/ng0893-332 · Cited by 1,340
Human housekeeping genes, revisited
Eisenberg, E., & Levanon, E. Y. (2013) · Trends in Genetics · doi:10.1016/j.tig.2013.05.010 · Cited by 1,249
Yates, A. D., Akanni, W. A., Amode, M. R., Barrell, D. G., Billis, K., Carvalho‐Silva, D., Cummins, C. A., Clapham, P., Fitzgerald, S. F., Gil, L., et al. (2015) · Nucleic Acids Research · doi:10.1093/nar/gkv1157 · Cited by 1,246
Streaming fragment assignment for real-time analysis of sequencing experiments
Roberts, A., & Pachter, L. (2013) · Nature Methods · doi:10.1038/nmeth.2251 · Cited by 1,063
De novo assembly and analysis of RNA-seq data
Robertson, G., Schein, J. E., Chiu, R., Corbett, R., Field, M. A., Jackman, S. D., Mungall, K. L., Lee, S. S., Okada, H. M., Qian, J. Q., et al. (2010) · Nature Methods · doi:10.1038/nmeth.1517 · Cited by 1,028
Hybrid error correction and de novo assembly of single-molecule sequencing reads
Koren, S., Schatz, M. C., Walenz, B. P., Martin, J. V., Howard, J. T., Ganapathy, G., Wang, Z., Rasko, D. A., McCombie, W. R., Jarvis, E. D., et al. (2012) · Nature Biotechnology · doi:10.1038/nbt.2280 · Cited by 1,025
SOAPdenovo-Trans: de novo transcriptome assembly with short RNA-Seq reads
Xie, Y., Wu, G., Tang, J., Luo, R., Patterson, J., Liu, S., Huang, W., He, G., Gu, S., Li, S., et al. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu077 · Cited by 1,012
SEQC/MAQC-III Consortium (2014) · Nature Biotechnology · doi:10.1038/nbt.2957 · Cited by 983 · cited 4×
“In terms of speed, STARlong was 68× faster than GMAP (Supplementary Table 8), while IDP took around 170 CPU hours to process each sample.”
MethodInfluentialHuang, W., Takebayashi, N., Qi, Y., & Hickerson, M. J. (2011) · BMC Bioinformatics · doi:10.1186/1471-2105-12-1 · Cited by 952
Assembling large genomes with single-molecule sequencing and locality-sensitive hashing
Berlin, K., Koren, S., Chin, C.-S., Drake, J. P., Landolin, J. M., & Phillippy, A. M. (2015) · Nature Biotechnology · doi:10.1038/nbt.3238 · Cited by 947
Ballgown bridges the gap between transcriptome assembly and expression analysis
Frazee, A. C., Pertea, G. M., Jaffe, A. E., Langmead, B., Salzberg, S. L., & Leek, J. T. (2015) · Nature Biotechnology · doi:10.1038/nbt.3172 · Cited by 866
LoRDEC: accurate and efficient long read error correction
Salmela, L., & Rivals, É. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu538 · Cited by 834
TopHat-Fusion: an algorithm for discovery of novel fusion transcripts
Kim, D., & Salzberg, S. L. (2011) · Genome biology · doi:10.1186/gb-2011-12-8-r72 · Cited by 816
Extensive sequencing of seven human genomes to characterize benchmark reference materials
Zook, J. M., Catoe, D., McDaniel, J., Vang, L., Spies, N., Sidow, A., Weng, Z., Liu, Y., Mason, C. E., Alexander, N., et al. (2016) · Scientific Data · doi:10.1038/sdata.2016.25 · Cited by 811 · cited 3×
“The accuracy was assessed on NA12878 using the NIST high-con fi dence (HC) calls 51 as the gold standard (Fig.”
MethodInfluentialSelf-citationThe HLA genomic loci map: expression, interaction, diversity and disease
Shiina, T., Hosomichi, K., Inoko, H., & Kulski, J. K. (2009) · Journal of Human Genetics · doi:10.1038/jhg.2008.5 · Cited by 795
Efficient de novo assembly of large genomes using compressed data structuresPreprint
Simpson, J. T., & Durbin, R. M. (2011) · Genome Research · doi:10.1101/gr.126953.111 · Cited by 767
Comprehensive evaluation of differential gene expression analysis methods for RNA-seq data
Rapaport, F., Khanin, R., Liang, Y., Pirun, M., Krek, A., Zumbo, P., Mason, C. E., Socci, N. D., & Betel, D. (2013) · Genome Biology · doi:10.1186/gb-2013-14-9-r95 · Cited by 733
A single-molecule long-read survey of the human transcriptome
Sharon, D., Tilgner, H., Grubert, F., & Snyder, M. (2013) · Nature Biotechnology · doi:10.1038/nbt.2705 · Cited by 701
Self-citationAssessment of transcript reconstruction methods for RNA-seq
The RGASP Consortium, Steijger, T., Abril, J. F., Engström, P. G., Kokocinski, F., Hubbard, T. J., Guigó, R., Harrow, J., & Bertone, P. (2013) · Nature Methods · doi:10.1038/nmeth.2714 · Cited by 699
RNA Editing Enzyme APOBEC1 and Some of Its Homologs Can Act as DNA Mutators
Harris, R. S., Petersen‐Mahrt, S. K., & Neuberger, M. S. (2002) · Molecular Cell · doi:10.1016/s1097-2765(02)00742-6 · Cited by 609
Patro, R., Mount, S. M., & Kingsford, C. (2014) · Nature Biotechnology · doi:10.1038/nbt.2862 · Cited by 586
Systematic evaluation of spliced alignment programs for RNA-seq data
The RGASP Consortium, Engström, P. G., Steijger, T., Sipos, B., Grant, G. R., Kahles, A., Rätsch, G., Goldman, N., Hubbard, T. J., Harrow, J., et al. (2013) · Nature Methods · doi:10.1038/nmeth.2722 · Cited by 513
proovread : large-scale high-accuracy PacBio correction through iterative short read consensus
Hackl, T., Hedrich, R., Schultz, J., & Förster, F. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu392 · Cited by 510
Gene expression in human embryonic stem cell lines: unique molecular signature
Bhattacharya, B., Miura, T., Brandenberger, R., Mejido, J., Luo, Y., Yang, A. X., Joshi, B. H., Ginis, I., Thies, R. S., Amit, M., et al. (2004) · Blood · doi:10.1182/blood-2003-09-3314 · Cited by 448
Widespread Polycistronic Transcripts in Fungi Revealed by Single-Molecule mRNA Sequencing
Gordon, S., Tseng, E., Salamov, A. A., Zhang, J., Meng, X., Zhao, Z., Kang, D., Underwood, J. G., Grigoriev, I. V., Figueroa, M., et al. (2015) · PLoS ONE · doi:10.1371/journal.pone.0132628 · Cited by 410
FusionCatcher – a tool for finding somatic fusion genes in paired-end RNA-sequencing dataPreprint
Nicorici, D., Şatalan, M., Edgren, H., Kangaspeska, S., Murumägi, A., Kallioniemi, O., Virtanen, S. M. M., & Kilkku, O. (2014) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/011650 · Cited by 406
Identifying RNA editing sites using RNA sequencing data alone
Ramaswami, G., Zhang, R., Piskol, R., Keegan, L. P., Deng, P., O’Connell, M. A., & Li, J. B. (2013) · Nature Methods · doi:10.1038/nmeth.2330 · Cited by 362
Accurate identification of A-to-I RNA editing in human by transcriptome sequencingPreprint
Bahn, J. H., Lee, J., Li, G., Greer, C., Peng, G., & Xiao, X. G. (2011) · Genome Research · doi:10.1101/gr.124107.111 · Cited by 352
Characterization of the human ESC transcriptome by hybrid sequencing
Au, K. F., Sebastiano, V., Afshar, P. T., Durruthy, J. D., Lee, L., Williams, B. A., van Bakel, H., Schadt, E. E., Reijo-Pera, R. A., Underwood, J. G., et al. (2013) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.1320101110 · Cited by 341
Self-citationImproving PacBio Long Read Accuracy by Short Read Alignment
Au, K. F., Underwood, J. G., Lee, L., & Wong, W. H. (2012) · PLoS ONE · doi:10.1371/journal.pone.0046679 · Cited by 335
Self-citationDefining a personal, allele-specific, and single-molecule long-read transcriptome
Tilgner, H., Grubert, F., Sharon, D., & Snyder, M. P. (2014) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.1400447111 · Cited by 266
Self-citationTilgner, H., Jahanbani, F., Blauwkamp, T., Moshrefi, A., Jaeger, E., Chen, F., Harel, I., Bustamante, C. D., Rasmussen, M., & Snyder, M. P. (2015) · Nature Biotechnology · doi:10.1038/nbt.3242 · Cited by 223
Self-citationSOAPfuse: an algorithm for identifying fusion transcripts from paired-end RNA-Seq data
Jia, W., Qiu, K., He, M., Song, P., Zhou, Q., Zhou, F., Yu, Y., Zhu, D., Nickerson, M. L., Wan, S., et al. (2013) · Genome biology · doi:10.1186/gb-2013-14-2-r12 · Cited by 206
A benchmark for RNA-seq quantification pipelines
Teng, M., Love, M. I., Davis, C., Djebali, S., Dobin, A., Graveley, B. R., Li, S., Mason, C. E., Olson, S., Pervouchine, D. D., et al. (2016) · Genome biology · doi:10.1186/s13059-016-0940-1 · Cited by 197
JAFFA: High sensitivity transcriptome-focused fusion gene detection
Davidson, N., Majewski, I. J., & Oshlack, A. (2015) · Genome Medicine · doi:10.1186/s13073-015-0167-x · Cited by 184
Han, H., & Li, X. (2011) · BMC Bioinformatics · doi:10.1186/1471-2105-12-s1-s7 · Cited by 170
Liu, S., Tsai, W.-H., Ding, Y., Chen, R., Fang, Z., Huo, Z., Kim, S., Ma, T., Chang, T., Priedigkeit, N., et al. (2015) · Nucleic Acids Research · doi:10.1093/nar/gkv1234 · Cited by 167
Genome sequence–independent identification of RNA editing sites
Zhang, Q., & Xiao, X. G. (2015) · Nature Methods · doi:10.1038/nmeth.3314 · Cited by 126
Weirather, J. L., Afshar, P. T., Clark, T. A., Tseng, E., Powers, L. S., Underwood, J. G., Zabner, J., Korlach, J., Wong, W. H., & Au, K. F. (2015) · Nucleic Acids Research · doi:10.1093/nar/gkv562 · Cited by 123
Self-citationSigma: Strain-level inference of genomes from metagenomic analysis for biosurveillance
Ahn, T.-H., Chai, J., & Pan, C. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu641 · Cited by 113
RapMap: a rapid, sensitive and accurate tool for mapping RNA-seq reads to transcriptomes
Srivastava, A., Sarkar, H., Gupta, N., & Patro, R. (2016) · Bioinformatics · doi:10.1093/bioinformatics/btw277 · Cited by 112
RNA-Seq Gene Profiling - A Systematic Empirical Comparison
Fonseca, N. A., Marioni, J. C., & Brāzma, A. (2014) · PLoS ONE · doi:10.1371/journal.pone.0107026 · Cited by 102
Fusion genes in solid tumors: an emerging target for cancer diagnosis and treatment
Parker, B. C., & Zhang, W. (2013) · 癌症:英文版 · doi:10.5732/cjc.013.10178 · Cited by 102
Hayer, K. E., Pizarro, A., Lahens, N. F., Hogenesch, J. B., & Grant, G. R. (2015) · Bioinformatics · doi:10.1093/bioinformatics/btv488 · Cited by 93 · cited 7×
“On MCF7-100 and MCF7-300 samples, Cuf fl inks-TopHat prediction sets were not enriched in any MCF7 or breast cancer-related gene expression study, while StringTie-HISAT2 ’ s and Salmon-SMEM ’ s top over-expressed genes were highly enriched in many MCF7 and breast cancer cell line-related gene sets (Supplementary Data 1…”
BackgroundMethodInfluentialMu, J. C., Mohiyuddin, M., Li, J., Asadi, N. B., Gerstein, M., Abyzov, A. S., Wong, W. H., & Lam, H. Y. K. (2014) · Bioinformatics · doi:10.1093/bioinformatics/btu828 · Cited by 76
Self-citationAccurate Identification and Analysis of Human mRNA Isoforms Using Deep Long Read Sequencing
Tilgner, H., Raha, D., Habegger, L., Mohiuddin, M., Gerstein, M., & Snyder, M. P. (2013) · G3 Genes Genomes Genetics · doi:10.1534/g3.112.004812 · Cited by 67
Self-citationGrape RNA-Seq analysis pipeline environment
Knowles, D. G., Röder, M., Merkel, A., & Guigó, R. (2013) · Bioinformatics · doi:10.1093/bioinformatics/btt016 · Cited by 34
RASER: reads aligner for SNPs and editing sites of RNA
Ahn, J., & Xiao, X. G. (2015) · Bioinformatics · doi:10.1093/bioinformatics/btv505 · Cited by 23
Brorsson, C., Hansen, N. T., Lage, K., Bergholdt, R., Brunak, S., & Pociot, F. (2009) · Diabetes Obesity and Metabolism · doi:10.1111/j.1463-1326.2008.01004.x · Cited by 21
Cited by
Sorted by citations
RNA sequencing: the teenage years
Stark, R., Grzelak, M., & Hadfield, J. (2019) · Nature Reviews Genetics · doi:10.1038/s41576-019-0150-2 · Cited by 2,041
RNA-Seq differential expression analysis: An extended review and a software tool
Silva, J. C., Domingues, D. S., & Lopes, F. M. (2017) · PLoS ONE · doi:10.1371/journal.pone.0190152 · Cited by 676
Souporcell: robust clustering of single-cell RNA-seq data by genotype without reference genotypes
Heaton, H., Talman, A. M., Knights, A., Imaz, M., Gaffney, D. J., Durbin, R., Hemberg, M., & Lawniczak, M. K. N. (2020) · Nature Methods · doi:10.1038/s41592-020-0820-1 · Cited by 506
Magic-BLAST, an accurate RNA-seq aligner for long and short reads
Boratyn, G. M., Thierry‐Mieg, J., Thierry‐Mieg, D., Busby, B., & Madden, T. (2019) · BMC Bioinformatics · doi:10.1186/s12859-019-2996-x · Cited by 345
Interpretation of differential gene expression results of RNA-seq data: review and integration
McDermaid, A., Monier, B., Zhao, J., Liu, B., & Ma, Q. (2018) · Briefings in Bioinformatics · doi:10.1093/bib/bby067 · Cited by 344
Metadata
- Authors
- Sahraeian, S. M. E., Mohiyuddin, M., Sebra, R., Tilgner, H., Afshar, P. T., Au, K. F., Bani Asadi, N., Gerstein, M. B., Wong, W. H., Snyder, M. P., Schadt, E., & Lam, H. Y. K.
- Affiliations
- Icahn School of Medicine at Mount Sinai (US)
- Stanford Medicine (US)
- Stanford University (US)
- University of Iowa (US)
- Yale University (US)
- Corresponding author
- Lam, Hugo Y. K.
- ORCID
- Venue
- Nature Communications
- Published
- 2017-07-05
- Volume
- 8
- Issue
- 1
- Pages
- 59-59
- ISSN
- 2041-1723 (ISSN-L)
- PMC
- PMC5498581
- OpenAlex
- W2731748842
- Cited by
- 376
- References
- 76
- Citations by year
- 2026: 29 · 2025: 48 · 2024: 64 · 2023: 41 · 2022: 49 · 2021: 60 · 2020: 28 · 2019: 31 · 2018: 20 · 2017: 5
- FWCI
- 9.38
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Locations
- Nature Communications (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- RePEc: Research Papers in Economics (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- Funders
- National Institutes of Health
- Genentech
- National Human Genome Research Institute (R01HG008759, P50HG007735, R01HG007834)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › RNA Research and Splicing
- Keywords
- gene expression profiling, RNA editing, computational pipeline, tumor transcriptome
- MeSH terms
- Base Sequence, Cell Line, Humans, Embryonic Stem Cells, Transcriptome
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
