

Protein sequence design by conformational landscape optimization
Open accessThe DOI matches its Crossref record; 39 of the 39 references it cites were found in a registry or an index.
Abstract
Significance: Almost all proteins fold to their lowest free energy state, which is determined by their amino acid sequence. Computational protein design has primarily focused on finding sequences that have very low energy in the target designed structure. However, what is most relevant during folding is not the absolute energy of the folded state but the energy difference between the folded state and the lowest-lying alternative states. We describe a deep learning approach that captures aspects of the folding landscape, in particular the presence of structures in alternative energy minima, and show that it can enhance current protein design methods.
Overview
The claims and statistics of this paper haven’t been checked yet.
Free: it reads the open-access text and checks the first 5 claims that cite a source we can check. Create a free account to check every claim.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Free: it reads the open-access text and checks the first 5 claims that cite a source we can check. Create a free account to check every claim.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
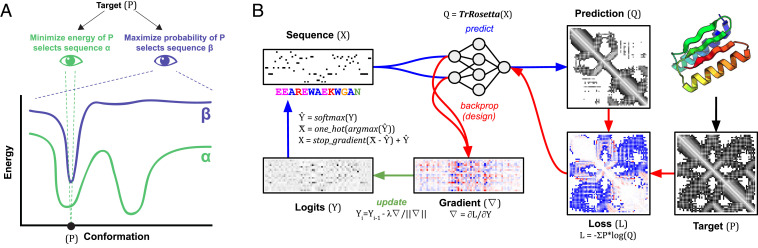
“trRosetta can also be used to design sequences that fold into a target backbone structure by carrying out sequence optimization using a structure recapitulation loss function that rewards similarity of the predicted structure to the target structure (13).”
From Scaffolding protein functional sites using deep learning (2022) · cites this paper as [13]
The check wasn’t confident either way, so it gives no verdict — never disputed.
In the citing paper
“We determined a 2.5 Å resolution crystal of L056 (see Methods, Figure 3e). The global fold was similar to predictions, with a Cα RMSD of 2.9 Å from the backbone structure predicted by trRosetta and 2.3 Å RMSD from a WT T4 Lysozyme structure59,60.”
From Large language models generate functional protein sequences across diverse families (2023) · cites this paper as [59]
Experimental — accuracy not yet measuredIn the citing paper
“RF and AF have different architectures and were trained independently, and hence AF predictions can be regarded as a partially orthogonal in silico test of whether RF-designed sequences fold into the intended structures, analogous to traditional ab initio folding (13, 24).”
From Scaffolding protein functional sites using deep learning (2022) · cites this paper as [13]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
A Simplex Method for Function Minimization
Nelder, J. A., & Mead, R. (1965) · The Computer Journal · doi:10.1093/comjnl/7.4.308 · Cited by 28,905
Amino acid substitution matrices from protein blocks.
Henikoff, S., & Henikoff, J. G. (1992) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.89.22.10915 · Cited by 6,218
TM-align: a protein structure alignment algorithm based on the TM-score
Zhang, Y. (2005) · Nucleic Acids Research · doi:10.1093/nar/gki524 · Cited by 3,774
Categorical Reparameterization with Gumbel-Softmax
Jang, E., Gu, S., & Poole, B. (2016) · arXiv · doi:10.48550/arxiv.1611.01144 · Cited by 3,616 · cited 1×
“3 with respect to the auxiliary variable Y (24):”
BackgroundHow fast-folding proteins fold
Lindorff-Larsen, K., Piana, S., Dror, R. O., & Shaw, D. E. (2011) · Science · doi:10.1126/science.1208351 · Cited by 1,980 · cited 1×
“…folding energy landscape characterization using molecular dynamics methods even for small proteins is a resource-intensive computational task (18, 19), and even Rosetta ab initio folding simulations are extremely computationally intensive, requiring thousands of CPU hours to generate enough…”
BackgroundAtomic-level characterization of the structural dynamics of proteins
Shaw, D. E., Maragakis, P., Lindorff-Larsen, K., Piana, S., Dror, R. O., Eastwood, M. P., Bank, J. A., Jumper, J. M., Salmon, J. K., Shan, Y., et al. (2010) · Science · doi:10.1126/science.1187409 · Cited by 1,923
Protein Structure Prediction Using Rosetta
Rohl, C. A., Strauss, C. E. M., Misura, K. M. S., & Baker, D. (2004) · Methods in Enzymology · doi:10.1016/s0076-6879(04)83004-0 · Cited by 1,725 · cited 1×
“Energy landscapes of designs were mapped out using Rosetta de novo structure prediction (ab initio folding) (27).”
MethodSelf-citationDesign of a Novel Globular Protein Fold with Atomic-Level Accuracy
Kuhlman, B., Dantas, G., Ireton, G. C., Varani, G., Stoddard, B. L., & Baker, D. (2003) · Science · doi:10.1126/science.1089427 · Cited by 1,670
Self-citationThe Rosetta All-Atom Energy Function for Macromolecular Modeling and Design
Alford, R. F., Leaver-Fay, A., Jeliazkov, J. R., O’Meara, M. J., DiMaio, F. P., Park, H., Shapovalov, M. V., Renfrew, P. D., Mulligan, V. K., Kappel, K., et al. (2017) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.7b00125 · Cited by 1,532
Self-citationImproved protein structure prediction using predicted interresidue orientations
Yang, J., Anishchenko, I., Park, H., Peng, Z., Ovchinnikov, S., & Baker, D. (2020) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.1914677117 · Cited by 1,421 · cited 2×
“Combining these predictions with Rosetta energy minimization yielded excellent predictions of structures in benchmark cases, as well as recent blind Continous Automated Model EvalutiOn (CAMEO) structure modeling evaluations (13).”
BackgroundMethodSelf-citationSimons, K. T., Kooperberg, C., Huang, E., & Baker, D. (1997) · Journal of Molecular Biology · doi:10.1006/jmbi.1997.0959 · Cited by 1,396 · cited 1×
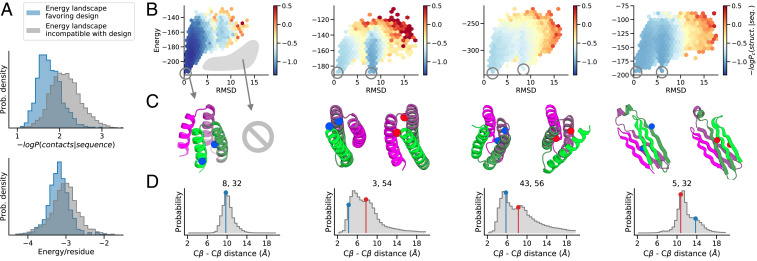
“Energy landscapes obtained from ab initio folding calculations are approximate as the sampling procedure is biased, nonexhaustive, and employs an imperfect energy function (17), but these biases should not qualitatively change our conclusions about overall landscape shape.”
Self-citationDe Novo Protein Design: Fully Automated Sequence Selection
Dahiyat, B. I., & Mayo, S. L. (1997) · Science · doi:10.1126/science.278.5335.82 · Cited by 1,145
Finite-time convergent gradient flows with applications to network consensus
Cortés, J. (2006) · Automatica · doi:10.1016/j.automatica.2006.06.015 · Cited by 818 · cited 1×
“where in order to havemore control over theminimization, we normalize the gradients (25) and gradually decrease the step size λi according to the nonlinear schedule λi = (1 − i=N)2, where N is the number of minimization steps.”
Generative models for graph-based protein design
Ingraham, J., Garg, V., Barzilay, R., & Jaakkola, T. (2019) · DSpace@MIT (Massachusetts Institute of Technology) · Cited by 662
RosettaScripts: A Scripting Language Interface to the Rosetta Macromolecular Modeling Suite
Fleishman, S. J., Leaver-Fay, A., Corn, J. E., Strauch, E.-M., Khare, S. D., Koga, N., Ashworth, J., Murphy, P., Richter, F., Lemmon, G., et al. (2011) · PLoS ONE · doi:10.1371/journal.pone.0020161 · Cited by 657 · cited 4×
“Backbones were constructed using blueprints (29) with distance and angle constraints to guide the formation of β -sheet backbone hydrogen bonds (30), followed by sequence design with Rosetta [FastDesign (28, 31) in conjunction with LayerDesign (21) and using the Rosetta all-atom energy function (32) with beta_nov16 we…”
BackgroundMethodInfluentialSelf-citationDe novo protein design by deep network hallucination
Anishchenko, I., Chidyausiku, T. M., Ovchinnikov, S., Pellock, S. J., & Baker, D. (2020) · doi:10.1101/2020.07.22.211482 · Cited by 629 · cited 1×
“We benchmarked this optimization method by redesigning 2,000 proteins previously generated using Markov chain Monte Carlo optimization with trRosetta (15).”
MethodSelf-citationPrinciples for designing ideal protein structures
Koga, N., Tatsumi-Koga, R., Liu, G., Xiao, R., Acton, T. B., Montelione, G. T., & Baker, D. (2012) · Nature · doi:10.1038/nature11600 · Cited by 564 · cited 2×
“While native structures usually require coevolution constraints derived from multiple sequence alignments to be predicted, the structures of de novo designed proteins, perhaps owing to their idealized sequence–structure encoding (9, 14), could be predicted from”
BackgroundSelf-citationGlobal analysis of protein folding using massively parallel design, synthesis, and testing
Rocklin, G. J., Chidyausiku, T. M., Goreshnik, I., Ford, A., Houliston, S., Lemak, A., Carter, L., Ravichandran, R., Mulligan, V. K., Chevalier, A., et al. (2017) · Science · doi:10.1126/science.aan0693 · Cited by 501 · cited 3×
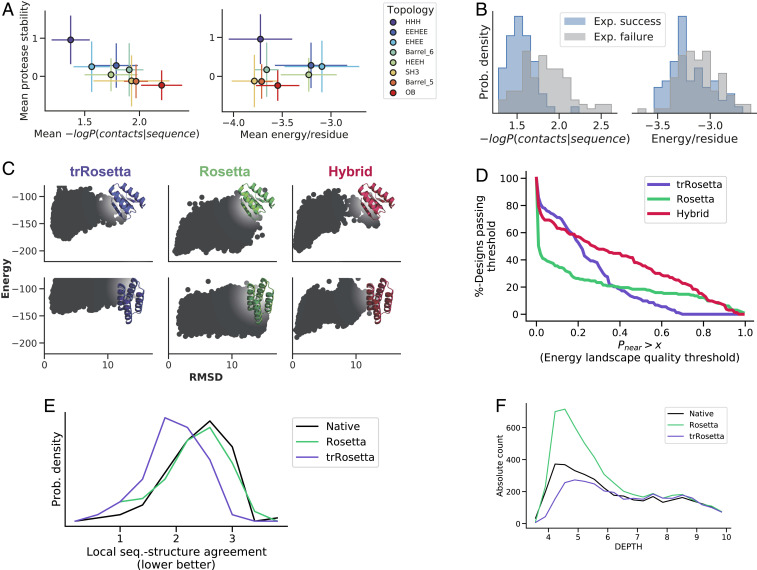
“Second, compared with Rosetta-based designs and native proteins, designs made with trRosetta have more ideal local sequence–structure relationships [an important determinant of design success (20, 23)] (Fig.”
MethodSelf-citationAccurate de novo design of hyperstable constrained peptides
Bhardwaj, G., Mulligan, V. K., Bahl, C. D., Gilmore, J. M., Harvey, P. J., Cheneval, O., Buchko, G. W., Pulavarti, S. V. S. R. K., Kaas, Q., Eletsky, A., et al. (2016) · Nature · doi:10.1038/nature19791 · Cited by 440 · cited 2×
“The quality of the energy landscape was quantified with Pnear (28), which approximates the Boltzmann-weighted probability of the structure adapting the target conformation (fuzzy cutoff):”
MethodSelf-citationRosettaRemodel: A Generalized Framework for Flexible Backbone Protein Design
Huang, P.-S., Ban, Y.-E. A., Richter, F., Andre, I., Vernon, R., Schief, W. R., & Baker, D. (2011) · PLoS ONE · doi:10.1371/journal.pone.0024109 · Cited by 369 · cited 1×
“Backbones were constructed using blueprints (29) with distance and angle constraints to guide the formation of β-sheet backbone hydrogen bonds (30), followed by sequence design with Rosetta [FastDesign (28, 31) in conjunction with LayerDesign (21) and using the Rosetta all-atom energy function (32) with beta_nov16 wei…”
MethodSelf-citationDe novo design of a fluorescence-activating β-barrel
Dou, J., Vorobieva, A. A., Sheffler, W., Doyle, L. A., Park, H., Bick, M. J., Mao, B., Foight, G. W., Lee, M. Y., Gagnon, L. A., et al. (2018) · Nature · doi:10.1038/s41586-018-0509-0 · Cited by 351 · cited 1×
“Backbones were constructed using blueprints (29) with distance and angle constraints to guide the formation of β-sheet backbone hydrogen bonds (30), followed by sequence design with Rosetta [FastDesign (28, 31) in conjunction with LayerDesign (21) and using the Rosetta all-atom energy function (32) with beta_nov16 wei…”
MethodSelf-citationAutomated design of specificity in molecular recognition
Havranek, J. J., & Harbury, P. B. (2003) · Nature Structural Biology · doi:10.1038/nsb877 · Cited by 309 · cited 1×
“Multistate design (10 – 12) can be carried out to maximize the energy gap between the desired conformation and other specified conformations, but the latter must be known in advance and be relatively few in number for such calculations to be tractable.”
BackgroundImproved protein structure refinement guided by deep learning based accuracy estimation
Hiranuma, N., Park, H., Baek, M., Anishchenko, I., Dauparas, J., & Baker, D. (2021) · Nature Communications · doi:10.1038/s41467-021-21511-x · Cited by 243
Self-citationProtein sequence design with a learned potential
Anand, N., Eguchi, R., Mathews, I. I., Perez, C. P., Derry, A., Altman, R. B., & Huang, P.-S. (2022) · Nature Communications · doi:10.1038/s41467-022-28313-9 · Cited by 171
Perturbing the energy landscape for improved packing during computational protein design
Maguire, J. B., Haddox, H. K., Strickland, D., Halabiya, S. F., Coventry, B., Griffin, J. R., Pulavarti, S. V. S. R. K., Cummins, M., Thieker, D. F., Klavins, E., et al. (2021) · Proteins: Structure, Function, and Bioinformatics · doi:10.1002/prot.26030 · Cited by 169
Self-citationDe novo protein design by citizen scientists
Koepnick, B., Flatten, J., Husain, T., Ford, A., Silva, D.-A., Bick, M. J., Bauer, A., Liu, G., Ishida, Y., Boykov, A., et al. (2019) · Nature · doi:10.1038/s41586-019-1274-4 · Cited by 160 · cited 4×
“To test this, we collected a set of 4,204 monomeric proteins designed by Foldit Players, who compete to optimize the Rosetta energy of designed structures without the aid of energy landscape calculations (16).”
InfluentialSelf-citationWatters, A. L., Deka, P., Corrent, C., Callender, D. R., Varani, G., Sosnick, T. R., & Baker, D. A. (2007) · Cell · doi:10.1016/j.cell.2006.12.042 · Cited by 156 · cited 1×
“A limitation of all of these approaches, however, is that while they generate a sequence that is the lowest-energy sequence for the desired structure, they can result in rough energy landscapes that hamper folding (7, 8) and do not guarantee that the desired structure is the lowest-energy structure for the sequence.”
BackgroundSelf-citationDesign of metalloproteins and novel protein folds using variational autoencoders
Greener, J. G., Moffat, L., & Jones, D. T. (2018) · Scientific Reports · doi:10.1038/s41598-018-34533-1 · Cited by 135
De novo protein design using pairwise potentials and a genetic algorithm
Jones, D. T. (1994) · Protein Science · doi:10.1002/pro.5560030405 · Cited by 130
DEPTH: a web server to compute depth and predict small-molecule binding cavities in proteins
Tan, K. P., Varadarajan, R., & Madhusudhan, M. S. (2011) · Nucleic Acids Research · doi:10.1093/nar/gkr356 · Cited by 98 · cited 1×
“The degree of burial was assessed with the software DEPTH (33), which computes the distance in ångströms between each residue and bulk solvent.”
MethodA Generic Program for Multistate Protein Design
Leaver-Fay, A., Jacak, R., Stranges, P. B., & Kuhlman, B. (2011) · PLoS ONE · doi:10.1371/journal.pone.0020937 · Cited by 94
An enumerative algorithm for de novo design of proteins with diverse pocket structures
Basanta, B., Bick, M. J., Bera, A. K., Norn, C., Chow, C. M., Carter, L. P., Goreshnik, I., Dimaio, F., & Baker, D. (2020) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.2005412117 · Cited by 82 · cited 1×
“Second, compared with Rosetta-based designs and native proteins, designs made with trRosetta have more ideal local sequence–structure relationships [an important determinant of design success (20, 23)] (Fig.”
MethodSelf-citationComputationally Designed Bispecific Antibodies using Negative State Repertoires
Leaver-Fay, A., Froning, K. J., Atwell, S., Aldaz, H., Pustilnik, A., Lu, F., Huang, F., Yuan, R., Hassanali, S., Chamberlain, A. K., et al. (2016) · Structure · doi:10.1016/j.str.2016.02.013 · Cited by 62 · cited 1×
“Multistate design (10 – 12) can be carried out to maximize the energy gap between the desired conformation and other specified conformations, but the latter must be known in advance and be relatively few in number for such calculations to be tractable.”
BackgroundFast activation maximization for molecular sequence design
Linder, J., & Seelig, G. (2021) · BMC Bioinformatics · doi:10.1186/s12859-021-04437-5 · Cited by 54
Protein Sequence Design with a Learned Potential
Anand-Achim, N., Eguchi, R. R., Mathews, I. I., Perez, C. P., Derry, A., Altman, R. B., & Huang, P.-S. (2020) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2020.01.06.895466 · Cited by 40 · cited 2×
“…landscape was quantified with P near (28), which approximates the Boltzmann-weighted probability of the structure adapting the target conformation (fuzzy cutoff): , [6] where N is the number of decoys, E is the energy of the decoy, and RMSD represents its structural deviation from the target state.”
BackgroundModular repeat protein sculpting using rigid helical junctions
Brunette, T., Bick, M. J., Hansen, J. M., Chow, C. M., Kollman, J. M., & Baker, D. (2020) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.1908768117 · Cited by 40 · cited 1×
“S11): LayerDesign, which disallows amino acids based on their surface exposure (21), and fragment-based PSSMs, which favor local sequence–structure relationships (22).”
BackgroundSelf-citationBasak, S., Nobrega, R. P., Tavella, D., Deveau, L. M., Koga, N., Tatsumi-Koga, R., Baker, D., Massi, F., & Matthews, C. R. (2019) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.1818744116 · Cited by 36 · cited 1×
“A limitation of all of these approaches, however, is that while they generate a sequence that is the lowest-energy sequence for the desired structure, they can result in rough energy landscapes that hamper folding (7, 8) and do not guarantee that the desired structure is the lowest-energy structure for the sequence.”
BackgroundSelf-citationTertiary Structural Propensities Reveal Fundamental Sequence/Structure Relationships
Zheng, F., Zhang, J., & Grigoryan, G. (2015) · Structure · doi:10.1016/j.str.2015.03.015 · Cited by 31 · cited 1×
“While native structures usually require coevolution constraints derived from multiple sequence alignments to be predicted, the structures of de novo designed proteins, perhaps owing to their idealized sequence–structure encoding (9, 14), could be predicted from”
BackgroundFast differentiable DNA and protein sequence optimization for molecular designPreprint
Linder, J., & Seelig, G. (2020) · arXiv (Cornell University) · doi:10.48550/arxiv.2005.11275 · Cited by 13
Cited by
Large language models generate functional protein sequences across diverse families
Madani, A., Krause, B., Greene, E. R., Subramanian, S., Mohr, B. P., Holton, J. M., Olmos, J. L., Xiong, C., Sun, Z. Z., Socher, R., et al. (2023) · Nature Biotechnology · doi:10.1038/s41587-022-01618-2 · Cited by 1,078
De novo protein design by deep network hallucination
Anishchenko, I., Chidyausiku, T. M., Ovchinnikov, S., Pellock, S. J., & Baker, D. (2020) · doi:10.1101/2020.07.22.211482 · Cited by 629
“On the sampling side, the Monte Carlo approach can be made more efficient by direct gradient-based minimization by tracing the gradients back to the inputs 25 .”
MethodSelf-citationAlphaFold2 and its applications in the fields of biology and medicine
Yang, Z., Zeng, X., Zhao, Y., & Chen, R. (2023) · Signal Transduction and Targeted Therapy · doi:10.1038/s41392-023-01381-z · Cited by 577
“111,112 The integrity of the structures predicted is validated and con fi rmed by standard ab initio folding, protein structure analysis methods, and rigorous all-atom molecular dynamics simulations.”
MethodScaffolding protein functional sites using deep learning
Wang, J., Lisanza, S., Juergens, D., Tischer, D., Watson, J. L., Castro, K. M., Ragotte, R., Saragovi, A., Milles, L. F., Baek, M., et al. (2022) · Science · doi:10.1126/science.abn2100 · Cited by 487
“…fold to the hallucinated structures (12). trRosetta can also be used to design sequences that fold into a target backbone structure by carrying out sequence optimization using a structure recapitulation loss function that rewards similarity of the predicted structure to the target structure (13).”
BackgroundMethodSelf-citationDe novo design of luciferases using deep learning
Yeh, A. H.-W., Norn, C., Kipnis, Y., Tischer, D., Pellock, S. J., Evans, D., Ma, P., Lee, G. R., Zhang, J. Z., Anishchenko, I., et al. (2023) · Nature · doi:10.1038/s41586-023-05696-3 · Cited by 441
“…of ideal NTF2-like structures, we developed a deep-learning-based ‘family-wide hallu-cination’ approach that integrates unconstrained de novo design 17,18 and Rosetta sequence-design approaches 19 to enable the generation of an essentially unlimited number of proteins that have a desired fold (Fig.”
MethodSelf-citation
Metadata
- Authors
- Norn, C., Wicky, B. I. M., Juergens, D., Liu, S., Kim, D., Tischer, D., Koepnick, B., Anishchenko, I., Players, F., Baker, D., Ovchinnikov, S., Coral, A., Bubar, A. J., Boykov, A., Valle Pérez, A. U., MacMillan, A., Lubow, A., Mussini, A., Cai, A., Ardill, A. J., Seal, A., Kalantarian, A., Failer, B., Lackersteen, B., Chagot, B., Haight, B. R., Taştan, B., Uitham, B., Roy, B. G., de Melo Cruz, B. R., Echols, B., Lorenz, B. E., Blair, B., Kestemont, B., Eastlake, C. D., Bragdon, C. J., Vardeman, C., Salerno, C., Comisky, C., Hayman, C. L., Landers, C. R., Zimov, C., Coleman, C. D., Painter, C. R., Ince, C., Lynagh, C., Malaniia, D., Wheeler, D. C., Robertson, D., Simon, V., Chisari, E., Kai, E. L. J., Rezae, F., Lengyel, F., Tabotta, F., Padelletti, F., Boström, F., Gross, G. O., McIlvaine, G., Beecher, G., Hansen, G. T., de Jong, G., Feldmann, H., Borman, J. L., Quinn, J., Norrgard, J., Truong, J., Diderich, J. A., Canfield, J. M., Photakis, J., Slone, J. D., Madzio, J., Mitchell, J., Stomieroski, J. C., Mitch, J. H., Altenbeck, J. R., Schinkler, J., Weinberg, J. B., Burbach, J. D., Sequeira da Costa, J. C., Bada Juarez, J. F., Gunnarsson, J. P., Harper, K. D., Joo, K., Clayton, K. T., DeFord, K. E., Scully, K. F., Gildea, K. M., Abbey, K. J., Kohli, K. L., Stenner, K., Takács, K., Poussaint, L. L., Manalo, L. C., Withers, L. C., Carlson, L., Wei, L., Fisher, L. R., Carpenter, L., Ji-hwan, M., Ricci, M., Belcastro, M. A., Leniec, M., Hohmann, M., Thompson, M., Thayer, M. A., Gaebel, M., Cassidy, M. D., Fagiola, M., Lewis, M., Pfützenreuter, M., Simon, M., Elmassry, M. M., Benevides, N., Kerr, N. K., Verma, N., Shannon, O., Yin, O., Wolfteich, P., Gummersall, P., Tłuścik, P., Gajar, P., Triggiani, P. J., Guha, R., Mathew Innes, R. B., Buchanan, R., Gamble, R., Leduc, R., Spearing, R., dos Santos Gomes, R. L. C., Estep, R. D., DeWitt, R., Moore, R., Shnider, S. G., Zaccanelli, S. J., Kuznetsov, S., Burillo-Sanz, S., Mooney, S., Vasiliy, S., Butkovich, S. S., Hudson, S. B., Pote, S. L., Denne, S. P., Schwegmann, S. A., Ratna, S., Kleinfelter, S. C., Bausewein, T., George, T. J., de Almeida, T. S., Yeginer, U., Barmettler, W., Pulley, W. R., Wright, W. S., Willyanto, Lansford, W., Hochart, X., Gaiji, Y. A. S., Lagodich, Y., & Christian, V.
- Affiliations
- University of Washington (US)
- Harvard University (US)
- Howard Hughes Medical Institute (US)
- Corresponding author
- Baker, David
- Ovchinnikov, Sergey
- ORCID
- Norn, Christoffer
- Wicky, Basile I. M.
- Juergens, David
- Liu, Sirui
- Kim, David
- Tischer, Doug
- Koepnick, Brian
- Anishchenko, Ivan
- Baker, David
- Ovchinnikov, Sergey
- Seal, Aniruddha
- Chagot, Benjamin
- Roy, Brandon G.
- Kestemont, Bruno
- Coleman, Charles David
- Chisari, Emanuele
- Truong, Jason
- Diderich, Jasper A.
- Slone, Jesse David
- Madzio, Joanna
- Joo, Keehyoung
- Wei, Linda
- Elmassry, Moamen M.
- Guha, Rajarshi
- Kuznetsov, Sergey
- Butkovich, Slava S.
- Pote, Spencer Len
- George, Thomas J.
- Venue
- Proceedings of the National Academy of Sciences
- Publisher
- National Academy of Sciences
- Published
- 2021-03-12
- Volume
- 118
- Issue
- 11
- ISSN
- 0027-8424 (ISSN-L)
- 1091-6490
- PMC
- PMC7980421
- OpenAlex
- W3134186384
- Cited by
- 144
- References
- 34
- Citations by year
- 2026: 19 · 2025: 15 · 2024: 26 · 2023: 33 · 2022: 30 · 2021: 21
- FWCI
- 8.62
- Type
- journal article
- Language
- en
- Open access
- hybrid
- doi.org
- Licence
- Locations
- Proceedings of the National Academy of Sciences (publishedVersion, cc-by-nc-nd)
- PubMed Central (publishedVersion, cc-by-nc-nd)
- PubMed (publishedVersion)
- Funders
- Novo Nordisk Foundation (NNF17OC0030446)
- European Molecular Biology Organization (ALTF 139-2018)
- HHS | NIH | NIH Office of the Director (DP5OD026389)
- Howard Hughes Medical Institute (N/A)
- Audacious Project at the Institute for Protein Design (N/A)
- HHS | NIH | National Institute of Allergy and Infectious Diseases (HHSN272201700059C)
- Schmidt Futures program (N/A)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Protein Structure and Dynamics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › RNA and protein synthesis mechanisms
- Physical Sciences › Chemistry › Organic Chemistry › Chemical Synthesis and Analysis
- Keywords
- protein design, backpropagation, de novo protein design, energy landscape, folding stability
- MeSH terms
- Models, Molecular, Protein Conformation, Proteins, Thermodynamics, Neural Networks, Computer, Protein Folding
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
Data sources
Bibliographic data: Semantic Scholar, OpenAlex, Crossref.
Contains information from the Semantic Scholar Academic Graph Datasets, made available under the ODC Attribution License (ODC-BY 1.0).
Semantic Scholar’s data is described in: Kinney, R., et al. (2023). The Semantic Scholar Open Data Platform. arXiv:2301.10140.
