

OpenMM 7: Rapid development of high performance algorithms for molecular dynamics
Open accessPublished version of a preprint
The DOI matches its Crossref record; 37 of the 37 references it cites were found in a registry or an index.
Abstract
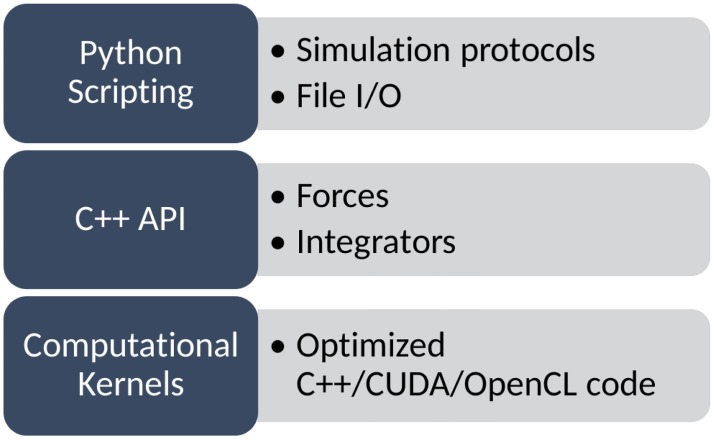
OpenMM is a molecular dynamics simulation toolkit with a unique focus on extensibility. It allows users to easily add new features, including forces with novel functional forms, new integration algorithms, and new simulation protocols. Those features automatically work on all supported hardware types (including both CPUs and GPUs) and perform well on all of them. In many cases they require minimal coding, just a mathematical description of the desired function. They also require no modification to OpenMM itself and can be distributed independently of OpenMM. This makes it an ideal tool for researchers developing new simulation methods, and also allows those new methods to be immediately available to the larger community.
Overview
The claims and statistics of this paper haven’t been checked yet.
Free: it reads the open-access text and checks the first 5 claims that cite a source we can check. Create a free account to check every claim.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Free: it reads the open-access text and checks the first 5 claims that cite a source we can check. Create a free account to check every claim.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“Possible other software engines to consider are ddcMD (Zhang et al., 2020) and openMM (Eastman et al., 2017), both of which are supporting Martini and offer simulation speeds comparable to those of Gromacs.”
From Molecular dynamics simulation of an entire cell (2023) · cites this paper as Eastman et al., 2017
Experimental — accuracy not yet measuredThe source backs part of the claim, not all of it — or backs it for a different population, condition or comparison.
In the citing paper
“The Boresch angles and dihedrals play the same role as the Euler and polar angles used in the geometrical route, but are easier to implement in other MD software, such as OpenMM.78”
From Assessment of the Topology and Oligomerisation States of Coiled Coils Using Metadynamics with Conformational Restraints (2025) · cites this paper as [78]
Experimental — accuracy not yet measured
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Comparison of simple potential functions for simulating liquid water
Jorgensen, W. L., Chandrasekhar, J., Madura, J. D., Impey, R. W., & Klein, M. L. (1983) · The Journal of Chemical Physics · doi:10.1063/1.445869 · Cited by 41,298 · cited 1×
“It loads a PDB file, models the forces with the AMBER99SB-ILDN force field [20] and TIP-3P water model [21], performs a local energy minimization to eliminate clashes, and then simulates 1 million steps of Verlet dynamics.”
MethodEquation of State Calculations by Fast Computing Machines
Metropolis, N., Rosenbluth, A. W., Rosenbluth, M. N., Teller, A. H., & Teller, E. (1953) · The Journal of Chemical Physics · doi:10.1063/1.1699114 · Cited by 37,192
Abraham, M. J., Murtola, T., Schulz, R., Páll, S., Smith, J. C., Hess, B., & Lindahl, E. (2015) · SoftwareX · doi:10.1016/j.softx.2015.06.001 · Cited by 27,190 · cited 12×
“For Gromacs and CHARMM, we compared only explicit solvent.”
MethodInfluentialCHARMM: The biomolecular simulation program
Brooks, B. R., Brooks, C. L., Mackerell, A. D., Nilsson, L., Petrella, R. J., Roux, B., Won, Y., Archontis, G., Bartels, C., Boresch, S., et al. (2009) · Journal of Computational Chemistry · doi:10.1002/jcc.21287 · Cited by 8,939 · cited 10×
“This includes the SWM4-NDP water model [34], and the CHARMM polarizable force field for proteins [4].”
MethodInfluentialSelf-citationImproved side‐chain torsion potentials for the Amber ff99SB protein force field
Lindorff‐Larsen, K., Piana, S., Palmo, K., Maragakis, P., Klepeis, J. L., Dror, R. O., & Shaw, D. E. (2010) · Proteins: Structure, Function, and Bioinformatics · doi:10.1002/prot.22711 · Cited by 6,134
Best, R. B., Zhu, X., Shim, J., Lopes, P. E. M., Mittal, J., Feig, M., & MacKerell, A. D. (2012) · Journal of Chemical Theory and Computation · doi:10.1021/ct300400x · Cited by 4,532 · cited 1×
“Some of the more important examples include the development of new force fields [1,2], sometimes involving novel functional forms for the interactions [3,4]; new integration algorithms [5–7]; new sampling methods [8–10]; and support for new types of hardware [11–13].”
BackgroundLee, J., Cheng, X., Swails, J. M., Yeom, M. S., Eastman, P. K., Lemkul, J. A., Wei, S., Buckner, J., Jeong, J. C., Qi, Y., et al. (2016) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.5b00935 · Cited by 4,171
Self-citationReversible multiple time scale molecular dynamics
Tuckerman, M., Berne, B. J., & Martyna, G. J. (1992) · The Journal of Chemical Physics · doi:10.1063/1.463137 · Cited by 3,592
Convergence acceleration of iterative sequences. the case of scf iteration
Pulay, P. (1980) · Chemical Physics Letters · doi:10.1016/0009-2614(80)80396-4 · Cited by 2,612 · cited 1×
“The iterative solver for induced dipoles uses the Direct Inversion in the Iterative Subspace (DIIS) algorithm [32] (added in OpenMM 6.”
MethodMolecular dynamics---Scalable algorithms for molecular dynamics simulations on commodity clusters
Bowers, K. J., Sacerdoti, F. D., Salmon, J. K., Shan, Y., Shaw, D. E., Chow, E., Xu, H., Dror, R. O., Eastwood, M. P., Gregersen, B. A., et al. (2006) · Proceedings of the 2006 ACM/IEEE conference on Supercomputing - SC '06 · doi:10.1145/1188455.1188544 · Cited by 2,580 · cited 3×
“OpenMM does include modelling tools, but it also can directly read the file formats used by Amber [16], CHARMM [17], Gromacs [18], and Desmond [19].”
MethodInfluential
Research works printed but not linked: 1
- Bekker H. Unification of box shapes in molecular simulations. J Comput Chem. 1997; 18: 1930– 1942. doi: 10.1002/(SICI)1096-987X(19971130)18:15<1930::AID-JCC8>3.0.CO;2-P · DOI
Cited by
Items 901–1,000 of 1,058
Benchmarking the Drude Polarizable Force Field Using the r(GACC) Tetranucleotide
Winkler, L., & Cheatham, T. E. (2023) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.3c00250 · Cited by 16
RNA-protein complexes and force field polarizability
Baltrukevich, H., & Bartos, P. (2023) · Frontiers in Chemistry · doi:10.3389/fchem.2023.1217506 · Cited by 16
“We want to note that the rather large default convergence threshold has been changed to 0.00001 in later versions of OpenMM (our simulations were run in early 2022).”
MethodInfluentialUnsaturated Lipids Facilitate Partitioning of Transmembrane Peptides into the Liquid Ordered Phase
Park, S., Levental, I., Pastor, R. W., & Im, W. (2023) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.3c00398 · Cited by 17
Application of computational approaches in biomembranes: From structure to function
Guo, J., Bao, Y., Li, M., Li, S., Xi, L., Xin, P., Wu, L., Liu, H., & Mu, Y. (2023) · Wiley Interdisciplinary Reviews Computational Molecular Science · doi:10.1002/wcms.1679 · Cited by 18
Variational deep learning of equilibrium transition path ensembles
Singh, A. N., & Limmer, D. T. (2023) · The Journal of Chemical Physics · doi:10.1063/5.0150278 · Cited by 16
Pandey, P., & MacKerell, A. D. (2023) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.3c00514 · Cited by 18
The inhibitory mechanism of a small protein reveals its role in antimicrobial peptide sensing
Jiang, S., Steup, L. C., Kippnich, C., Lazaridi, S., Malengo, G., Lemmin, T., & Yuan, J. (2023) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.2309607120 · Cited by 18
MiMiC: A high-performance framework for multiscale molecular dynamics simulations
Antalík, A., Levy, A., Kvedaravičiūtė, S., Johnson, S. K., Carrasco‐Busturia, D., Raghavan, B., Mouvet, F., Acocella, A., Das, S. K., Gavini, V., et al. (2024) · The Journal of Chemical Physics · doi:10.1063/5.0211053 · Cited by 16
Hentrich, C., Putyrski, M., Hanuschka, H., Preis, W., Kellmann, S.-J., Wich, M., Cavada, M., Hanselka, S., Lelyveld, V. S., & Ylera, F. (2024) · Nature Communications · doi:10.1038/s41467-024-50296-y · Cited by 16
“The resulting base model was then mutated to introduce each of eight possible biotinylated cysteine mutants I27C, F29C, A42C, M44C, S59C, F75C, Y84C, and F92C using PDBFixer 43 .”
Devasani, J. R., Guntuku, G., & Palla, M. S. (2024) · Biotechnology and Bioengineering · doi:10.1002/bit.28816 · Cited by 16
García‐Rivas, G., Lozano, O., Bernal‐Ramírez, J., Silva‐Platas, C., Salazar‐Ramírez, F., Méndez‐Fernández, A., Morales‐Ochoa, C. A., Alves‐Figueiredo, H., Ramos-González, M., Rubio‐Infante, N., et al. (2025) · JACC Basic to Translational Science · doi:10.1016/j.jacbts.2024.12.009 · Cited by 21
ELF: An Extended-Lagrangian Free Energy Calculation Module for Multiple Molecular Dynamics Engines
Chen, H., Fu, H., Shao, X., Chipot, C. J., & Cai, W. (2018) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.8b00115 · Cited by 16
Janus: An Extensible Open-Source Software Package for Adaptive QM/MM Methods
Zhang, B., Altarawy, D., Barnes, T., Turney, J. M., & Schaefer, H. F. (2019) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.9b00182 · Cited by 16
Zhang, J., & Gong, H. (2020) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.0c00064 · Cited by 16
Computing Ligands Bound to Proteins Using MELD-Accelerated MD
Liu, C., Brini, E., Pérez, A., & Dill, K. A. (2020) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.0c00543 · Cited by 17
“42 Two types of information are input into MELD as restraints.”
BackgroundCyclic Peptides as Protein Kinase Inhibitors: Structure–Activity Relationship and Molecular Modeling
Sanner, M. F., Zoghebi, K. A., Hanna, S. E., Mozaffari, S., Rahighi, S., Tiwari, R. K., & Parang, K. (2021) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.1c00320 · Cited by 16
Vilseck, J. Z., Ding, X., Hayes, R. L., & Brooks, C. L. (2021) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.1c00176 · Cited by 17
Progress in Free Energy Perturbation: Options for Evolving Fragments
Zara, L., Efrém, N., van Muijlwijk‐Koezen, J. E., de Esch, I. J. P., & Zarzycka, B. A. (2021) · Drug Discovery Today Technologies · doi:10.1016/j.ddtec.2021.10.001 · Cited by 16
Wang, S., Krummenacher, K., Landrum, G. A., Sellers, B. D., Lello, P. D., Robinson, S. J., Martin, B. T., Holden, J. K., Tom, J., Murthy, A. C., et al. (2022) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.1c01165 · Cited by 16
Mahmoud, A. H., Masters, M. R., Lee, S. J., & Lill, M. A. (2022) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.1c01438 · Cited by 20
Obtaining QM/MM binding free energies in the SAMPL8 drugs of abuse challenge: indirect approaches
Hudson, P. S., Aviat, F., Meana‐Pañeda, R., Warrensford, L. C., Pollard, B. C., Prasad, S., Jones, M. R., Woodcock, H. L., & Brooks, B. R. (2022) · Journal of Computer-Aided Molecular Design · doi:10.1007/s10822-022-00443-8 · Cited by 17
Self-citationNaseem, M. U., Carcamo-Noriega, E. N., Beltrán-Vidal, J., Borrego, J., Szanto, T. G., Zamudio, F. Z., Delgado-Prudencio, G., Possani, L. D., & Panyi, G. (2022) · The Journal of General Physiology · doi:10.1085/jgp.202213146 · Cited by 17
de Lima, T. A. D. M., de Lima, G. G., Chee, B. S., Henn, J. G., Cortese, Y. J., Matos, M., Helm, C. V., Magalhães, W. L. E., & Nugent, M. (2022) · Polymers · doi:10.3390/polym14142776 · Cited by 18
“(2021) [28] which utilizes the google collab feature to run the google compute engine backend for this simulation using the OpenMM molecular dynamics code [29].”
MethodTrifan, A., Gorgun, D., Salim, M., Brace, A., Zvyagin, M., Ma, H., Clyde, A., Clark, D., Salim, M., Hardy, D. J., et al. (2022) · The International Journal of High Performance Computing Applications · doi:10.1177/10943420221113513 · Cited by 16
“Missing loop residues ARG ALA ARG corresponding to residue numbers 314-316 in the nsp13 subunits were modeled using PDBFIXER tool in OpenMM (Eastman et al., 2017).”
MethodA thermodynamic investigation of amyloid precursor protein processing by human γ-secretase
Lu, X., & Huang, J. (2022) · Communications Biology · doi:10.1038/s42003-022-03818-7 · Cited by 16
An open-source molecular builder and free energy preparation workflow
Bieniek, M., Cree, B., Pirie, R., Horton, J. T., Tatum, N. J., & Cole, D. J. (2022) · Communications Chemistry · doi:10.1038/s42004-022-00754-9 · Cited by 16
“The protein is treated with PDBFixer 38 to add any missing atoms, residues, and hydrogen atoms.”
BackgroundMethodPyMM: An Open-Source Python Program for QM/MM Simulations Based on the Perturbed Matrix Method
Chen, C. G., Nardi, A. N., Amadei, A., & D’Abramo, M. (2022) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.2c00767 · Cited by 16
Charge transfer as a mechanism for chlorophyll fluorescence concentration quenching
Worster, S. B., Feighan, O., & Manby, F. R. (2023) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.2210811120 · Cited by 16
“OpenMM (37) for 12 different systems: a pair of neutral chlorophyll molecules (Chl – Chl) or a pair of chlorophyll ions (Chl+ – Chl) embedded in light-”
BackgroundBallard, C. J., Paserba, M., Daniel, E. J. P., Hurtado‐Guerrero, R., & Gerken, T. A. (2023) · Glycobiology · doi:10.1093/glycob/cwad066 · Cited by 16
Khuttan, S., Azimi, S., Wu, J. Z., Dick, S., Wu, C., Xu, H., & Gallicchio, E. (2023) · Physical Chemistry Chemical Physics · doi:10.1039/d3cp02125d · Cited by 18
“Reaching convergence for some complexes involving slow intramolecular degrees of freedom required advanced metadynamicsbased conformational sampling strategies,[35, 36] which”
BackgroundMethodCost-Effective Potential for Accurate Polarizable Embedding Calculations in Protein Environments
Reinholdt, P., Kjellgren, E. R., Steinmann, C., & Olsen, J. M. H. (2019) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.9b00616 · Cited by 16
Yang, D. T., Gronenborn, A. M., & Chong, L. T. (2022) · The Journal of Physical Chemistry A · doi:10.1021/acs.jpca.2c00255 · Cited by 17
Improving Antibody Design with Force-Guided Sampling in Diffusion ModelsPreprint
Kulytė, P., Vargas, F., Mathis, S. V., Wang, Y. G., Hernández-Lobato, J. M., & Lió, P. (2024) · arXiv (Cornell University) · doi:10.48550/arxiv.2406.05832 · Cited by 22
“Finally, the generated structures, as well as reference original ones, are relaxed using OpenMM (Eastman et al., 2017) and Rosetta (Alford et al., 2017).”
How Robust Is the Ligand Binding Transition State?
Bose, S., Lotz, S. D., Deb, I., Shuck, M., Lee, K. S. S., & Dickson, A. (2023) · Journal of the American Chemical Society · doi:10.1021/jacs.3c08940 · Cited by 16
An in-depth review of AI-powered advancements in cancer drug discovery
Le, M. H. N., Nguyen, P. K., Nguyen, T. P. T., Nguyen, H. Q., Tam, D. N. H., Huynh, H. H., Huynh, P. K., & Le, N. Q. K. (2025) · Biochimica et Biophysica Acta (BBA) - Molecular Basis of Disease · doi:10.1016/j.bbadis.2025.167680 · Cited by 15
Structural reorganization and relaxation dynamics of axially stressed chromosomes
Ruben, B. S., Brahmachari, S., Contessoto, V. G., Cheng, R. R., Junior, A. B. D. O., Pierro, M. D., & Onuchic, J. N. (2023) · Biophysical Journal · doi:10.1016/j.bpj.2023.03.029 · Cited by 16
Retamal-Farfán, I., González-Higueras, J., Galaz‐Davison, P., Rivera, M., & Ramírez‐Sarmiento, C. A. (2023) · Biophysical Reviews · doi:10.1007/s12551-023-01087-0 · Cited by 14
Structural Modeling of Cytokine-Receptor-JAK2 Signaling Complexes Using AlphaFold Multimer
Pogozheva, I. D., Cherepanov, S., Park, S., Raghavan, M., Im, W., & Lomize, A. L. (2023) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.3c00926 · Cited by 19
Yang, Y., Liu, Y., Zhang, J., Yang, Q., Zhang, J., Chu, H., Zhao, H., Luo, M., Lu, X., Liu, D., et al. (2023) · PLoS Biology · doi:10.1371/journal.pbio.3002285 · Cited by 15
“For each ligand, the top 1 docking model was selected and energy minimization was performed by OpenMM7 [35] with the Amber14 [36] force field to optimize the complex model.”
MethodPolonius, S., Zhuravel, O., Bachmair, B., & Mai, S. (2023) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.3c00805 · Cited by 16
Guilvout, I., Samsudin, F., Huber, R. G., Bond, P. J., Bardiaux, B., & Francetić, O. (2023) · mBio · doi:10.1128/mbio.01423-23 · Cited by 16
Qandeel, B. M., Mowafy, S., Abouzid, K. A. M., & Farag, N. A. (2024) · BMC Chemistry · doi:10.1186/s13065-023-01110-1 · Cited by 16
“All simulations in this work were done using the GPU-accelerated version of OpenMM 7.6 [81] engine and the ‘Making it rain’ [82] cloud-based molecular simulations notebook environment.”
MethodInfluentialFasoulis, R., Rigo, M., Lizee, G. A., Antunes, D. A., & Kavraki, L. E. (2024) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.3c01667 · Cited by 18
Bai, H., Lewitus, É., Li, Y., Thomas, P. V., Zemil, M., Merbah, M., Peterson, C. E., Thuraisamy, T., Rees, P. A., Hajduczki, A., et al. (2024) · Nature Communications · doi:10.1038/s41467-024-48139-x · Cited by 15
Olehnovics, E., Liu, Y. M., Mehio, N., Sheikh, A. Y., Shirts, M. R., & Salvalaglio, M. (2024) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.4c00520 · Cited by 15
Dynamic Ion Correlations and Ion-Pair Lifetimes in Aqueous Alkali Metal Chloride Electrolytes
Elliott, G. R., Wanless, E. J., Webber, G. B., Andersson, G. G., Craig, V. S. J., & Page, A. J. (2024) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.4c01992 · Cited by 16
Systematic Approach to Parametrization of Disaccharides for the Martini 3 Coarse-Grained Force Field
Brandner, A. F., Smith, I. P. S., Marrink, . J., Souza, P. C. T., & Khalid, S. (2025) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.4c01874 · Cited by 15
Large-scale computational modelling of H5 influenza variants against HA1-neutralising antibodies
Ford, C. T., Yasa, S., Obeid, K., Jaimes, R., Tomezsko, P. J., Guirales-Medrano, S., White, R. A., III, & Janies, D. A. (2025) · EBioMedicine · doi:10.1016/j.ebiom.2025.105632 · Cited by 15
High Accuracy Protein Structures from Minimal Sparse Paramagnetic Solid‐State NMR Restraints
Pérez, A., Gaalswyk, K., Jaroniec, C. P., & MacCallum, J. L. (2019) · Angewandte Chemie International Edition · doi:10.1002/anie.201811895 · Cited by 16
Chain-Increment Method for Free-Energy Computation of a Polymer with All-Atom Molecular Simulations
Yamada, K., & Matubayasi, N. (2020) · Macromolecules · doi:10.1021/acs.macromol.9b01952 · Cited by 15
Quantifying Polypeptoid Conformational Landscapes through Integrated Experiment and Simulation
Jiao, S., DeStefano, A. J., Monroe, J. I., Barry, M. E., Sherck, N. J., Casey, T. M., Segalman, R. A., Han, S., & Shell, M. S. (2021) · Macromolecules · doi:10.1021/acs.macromol.1c00550 · Cited by 16
García‐Marín, J., Griera, M., Alajarı́n, R., Rodrı́guez-Puyol, M., Rodrı́guez-Puyol, D., & Vaquero, J. J. (2021) · ChemMedChem · doi:10.1002/cmdc.202100338 · Cited by 15
Structural-Based Modeling in Protein Engineering. A Must Do
Rodà, S., Robles‐Martín, A., Xiang, R., Kazemi, M., & Guallar, V. (2021) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.1c02545 · Cited by 17
Sengul, M. Y., & MacKerell, A. D. (2021) · Journal of Computational Biophysics and Chemistry · doi:10.1142/s2737416521420060 · Cited by 15
Konc, J., Lešnik, S., Škrlj, B., Sova, M., Proj, M., Knez, D., Gobec, S., & Janežič, D. (2022) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.1c01176 · Cited by 15
McDaniel, J. G. (2022) · The Journal of Physical Chemistry C · doi:10.1021/acs.jpcc.2c00256 · Cited by 16
Sydow, D., Rodríguez-Guerra, J., Kimber, T. B., Schaller, D., Taylor, C. J., Chen, Y., Leja, M., Misra, S., Wichmann, M., Ariamajd, A., et al. (2022) · Nucleic Acids Research · doi:10.1093/nar/gkac267 · Cited by 17
“(46) for further reading on different MD protocols run with OpenMM us-ing Jupyter notebooks on Google Colab.”
MethodSpinaci, A., Buccioni, M., Catarzi, D., Cui, C., Colotta, V., Ben, D. D., Cescon, E., Francucci, B., Grieco, I., Lambertucci, C., et al. (2023) · Pharmaceuticals · doi:10.3390/ph16020167 · Cited by 16
“The code to run SuMD simulations is written in the Python programming language and exploits the ACEMD3 engine [63], which is based on the OpenMM7 Python library [64], to perform molecular dynamics simulations, while the geometric supervision of the trajectories is carried out through the ProDy library [80].”
MethodInfluentialKatzberger, P., & Riniker, S. (2023) · The Journal of Chemical Physics · doi:10.1063/5.0147027 · Cited by 18
Bi, J., Thiel, K. W., Litman, J. M., Zhang, Y., Devor, E. J., Newtson, A. M., Schnieders, M. J., Bosquet, J. G., & Leslie, K. K. (2019) · Clinical Obstetrics & Gynecology · doi:10.1097/grf.0000000000000516 · Cited by 15
Kraus, J., Gupta, R., Lu, M., Gronenborn, A. M., Akke, M., & Polenova, T. E. (2020) · ChemPhysChem · doi:10.1002/cphc.202000249 · Cited by 15
Amin, K. S., Hu, X., Salahub, D. R., Baldauf, C., Lim, C., & Noskov, S. Y. (2020) · The Journal of Chemical Physics · doi:10.1063/5.0020768 · Cited by 15
Chen, H., Deng, J., Cui, Q., Chanda, B., & Henzler‐Wildman, K. A. (2021) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.2017280118 · Cited by 15
“J. Lee et al., CHARMM-GUI input generator for NAMD, GROMACS, AMBER, OpenMM, and CHARMM/OpenMM simulations using the CHARMM36 additive force field.”
MethodInfluentialOpen Science Discovery of Oral Non-Covalent SARS-CoV-2 Main Protease InhibitorsPreprint
The COVID Moonshot Consortium, Chodera, J. D., Lee, A. A., London, N., & von Delft, F. (2021) · ChemRxiv · doi:10.26434/chemrxiv-2021-585ks-v2 · Cited by 15
Self-citationRNA encodes physical informationPreprint
Seim, I., Zhang, V., Jalihal, A. P., Stormo, B. M., Cole, S. J., Ekena, J., Nguyen, H. T., Thirumalai, D., & Gladfelter, A. S. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.12.11.627970 · Cited by 19
Mind the Gap—Deciphering GPCR Pharmacology Using 3D Pharmacophores and Artificial Intelligence
(2022) · doi:10.3390/ph15111304 · Cited by 15
“The extensive open-source software OpenMM [127] provides an MD simulation toolkit, including a tool for the preparation of the molecular system, which allows free development of new simulation protocols or novel functional forms of interactions.”
MethodQuantification of the Impact of Structure Quality on Predicted Binding Free Energy Accuracy
Behera, S., Hahn, D. F., Wilson, C. J., Marsili, S., Tresadern, G., Gapsys, V., & de Groot, B. L. (2025) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.5c00947 · Cited by 14
Two CTCF motifs impede cohesin-mediated DNA loop extrusion
Barth, R., Janissen, R., Muras, L., van der Torre, J., Litos, G., van der Sluis, E. O., van den Berg, A., Davidson, I. F., Peters, J., & Dekker, C. (2025) · Molecular Cell · doi:10.1016/j.molcel.2025.11.001 · Cited by 20
Rosignoli, S., Lustrino, E., Shevchuk, O., Rinaldo, S., Rubini, E., Paiardini, A., & Carev, I. (2026) · Biomolecules · doi:10.3390/biom16020197 · Cited by 16
“Molecular Dynamics GROMACS [196], OpenMM [233], NAMD [234], CHARMM-GUI [235], AmberTools [236] MD simulation software 6.”
The filamentous ultrastructure of the PopZ condensate is required for its cellular function
Scholl, D., Boyd, T., Latham, A. P., Salazar, A., Khan, A. M. A. M., Boeynaems, S., Holehouse, A. S., Lander, G. C., Sali, A., Park, D., et al. (2026) · Nature Structural & Molecular Biology · doi:10.1038/s41594-025-01742-y · Cited by 14
“Simulations were performed in OpenMM (version 7.5.1) 82 , using the OpenABC software package (version 1.06) 83 .”
InfluentialTeng, X., Liu, B., & Ichiye, T. (2020) · The Journal of Chemical Physics · doi:10.1063/5.0021472 · Cited by 14
Raybould, M. I. J., Nissley, D. A., Kumar, S., & Deane, C. M. (2023) · Frontiers in Immunology · doi:10.3389/fimmu.2022.1080596 · Cited by 16
Understanding FABP7 binding to fatty acid micelles and membranes
Lenz, S. A. P., Bodnariuc, I., Renaud-Young, M. M., Butler, T. M., & MacCallum, J. L. (2023) · Biophysical Journal · doi:10.1016/j.bpj.2023.01.023 · Cited by 15
Assessments of Variational Autoencoder in Protein Conformation Exploration
Xiao, S., Song, Z., Tian, H., & Tao, P. (2023) · Journal of Computational Biophysics and Chemistry · doi:10.1142/s2737416523500217 · Cited by 16
Heparan sulfate glycomimetics via iterative assembly of “clickable” disaccharides
Yang, C., Deng, Y., Wang, Y., Xia, C., Mehta, A. Y., Baker, K. J., Samal, A., Booneimsri, P., Lertmaneedang, C., Hwang, S. H., et al. (2023) · Chemical Science · doi:10.1039/d3sc00260h · Cited by 15
Peng: A program for transport properties of low-density binary gas mixtures
Zhai, Y., Li, Y., Li, H., & McCourt, F. R. W. (2023) · Computer Physics Communications · doi:10.1016/j.cpc.2023.108712 · Cited by 14
“objects (or things) in real life, it is a common practice [13, 14] to present an algorithm as an object.”
BackgroundUracil-DNA glycosylase efficiency is modulated by substrate rigidity
Orndorff, P. B., Poddar, S., Owens, A. M., Kumari, N., Ugaz, B. T., Amin, S. A., Horn, W. D. V., van der Vaart, A., & Levitus, M. (2023) · Scientific Reports · doi:10.1038/s41598-023-30620-0 · Cited by 16
Morado, J., Mortenson, P. N., Nissink, J. W. M., Essex, J. W., & Skylaris, C. (2023) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.2c01510 · Cited by 18
“All MD simulations were run in OpenMM, 95 and those that used ML models used the openmm-ml plugin that can be found at https://github.com/openmm/openmm-ml.”
Kehrein, J., & Sotriffer, C. (2023) · ACS Biomaterials Science & Engineering · doi:10.1021/acsbiomaterials.3c00636 · Cited by 14
Dohn, A. O., Markmann, V. I., Nimmrich, A., Haldrup, K., Møller, K. B., & Nielsen, M. M. (2023) · The Journal of Chemical Physics · doi:10.1063/5.0164365 · Cited by 14
Rubina, R., & Moin, S. T. (2023) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.3c01130 · Cited by 14
Improved Sampling of Adaptive Path Collective Variables by Stabilized Extended-System Dynamics
Hulm, A., & Ochsenfeld, C. (2023) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.3c00938 · Cited by 15
A Fast, Convenient, Polarizable Electrostatic Model for Molecular Dynamics
Wang, L., Schauperl, M., Mobley, D. L., Bayly, C. I., & Gilson, M. K. (2024) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.3c01171 · Cited by 14
Self-citationAbFlex: designing antibody complementarity determining regions with flexible CDR definition
Jeon, W., & Kim, D. (2024) · Bioinformatics · doi:10.1093/bioinformatics/btae122 · Cited by 14
“Next, we reconstruct the rest of the CDR atoms, including backbone and side chain atoms, using PDBFixer (Eastman et al. 2017).”
MethodInfluentialCiancetta, A., Malfacini, D., Gozzi, M., Marzola, E., Camilotto, R., Calò, G., & Guerrini, R. (2024) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.4c00499 · Cited by 17
Alsharabasy, A. M., Lagarias, P. I., Papavasileiou, K. D., Afantitis, A., Farràs, P., Glynn, S. A., & Pandit, A. S. (2024) · Journal of Medicinal Chemistry · doi:10.1021/acs.jmedchem.4c00989 · Cited by 15
Kihn, K. C., Purdy, O., Lowe, V. M., Šlachtová, L., Smith, A. K., Shapiro, P., & Deredge, D. J. (2024) · Journal of the American Society for Mass Spectrometry · doi:10.1021/jasms.4c00202 · Cited by 15
Vornweg, J. R., & Jacob, C. R. (2024) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.4c05645 · Cited by 14
Leonardo, J., Putra, P. P., Tallei, T. E., Fatimawali, F., Taslim, N. A., Tjandrawinata, R. R., Lau, V., Mustafa, S., Prieto, M. A., & Nurkolis, F. (2024) · Pharmacological Research - Modern Chinese Medicine · doi:10.1016/j.prmcm.2024.100551 · Cited by 14
Evolutionary Dynamics of RuBisCO: Emergence of the Small Subunit and its Impact Through Time
Amritkar, K., Cuevas-Zuviría, B., & Kaçar, B. (2024) · Molecular Biology and Evolution · doi:10.1093/molbev/msae268 · Cited by 16
“We carried out the MD simulation on OpenMM 7.7 (Eastman et al. 2017) running in the Nvidia Tesla A100, L40 and H100 GPU nodes of the Center for High Throughput Computation at the University of Wisconsin-Madison.”
InfluentialSetiadi, J., Biedermann, F., Nau, W. M., & Gilson, M. K. (2025) · Angewandte Chemie International Edition · doi:10.1002/anie.202505713 · Cited by 14
Correlation between Labeling Yield and Surface Accessibility in Covalent Labeling Mass Spectrometry
Ziemianowicz, D. S., MacCallum, J. L., & Schriemer, D. C. (2020) · Journal of the American Society for Mass Spectrometry · doi:10.1021/jasms.9b00083 · Cited by 14
Proton Transport in [BMIM+][BF4–]/Water Mixtures Near the Percolation Threshold
Stoppelman, J. P., & McDaniel, J. G. (2020) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.0c02487 · Cited by 15
Proton Transfer Dynamics in the Aprotic Proton Accepting Solvent 1-Methylimidazole
Thomaz, J. E., Walker, A. R., Wyck, S. J. V., Meisner, J., Martı́nez, T. J., & Fayer, M. D. (2020) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.0c05525 · Cited by 14
Jung, H., & Yethiraj, A. (2020) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.0c06510 · Cited by 14
Juárez‐Jiménez, J., Tew, P., O’Connor, M. P., Llabrés, S., Sage, R., Glowacki, D. R., & Michel, J. (2020) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.0c00221 · Cited by 15
Current computational methods for enzyme design
Coates, T. L., Young, N. J., Jarrett, A. J., Morris, C. J., Moody, J., & Corte, D. D. (2021) · Modern Physics Letters B · doi:10.1142/s0217984921501554 · Cited by 14
Influences of Vibrio cholerae Lipid A Types on LPS Bilayer Properties
Luna, E., Kim, S., Gao, Y., Widmalm, G., & Im, W. (2021) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.0c09144 · Cited by 14
Continuous Constant pH Molecular Dynamics Simulations of Transmembrane Proteins
Huang, Y., Henderson, J. A., & Shen, J. (2021) · Methods in molecular biology · doi:10.1007/978-1-0716-1394-8_15 · Cited by 14
Song, Z., Trozzi, F., Tian, H., Yin, C., & Tao, P. (2022) · ACS Physical Chemistry Au · doi:10.1021/acsphyschemau.2c00005 · Cited by 14
Metadata
- Authors
- Eastman, P., Swails, J., Chodera, J. D., McGibbon, R. T., Zhao, Y., Beauchamp, K. A., Wang, L.-P., Simmonett, A. C., Harrigan, M. P., Stern, C. D., Wiewiora, R. P., Brooks, B. R., & Pande, V. S.
- Affiliations
- Stanford University (US)
- Rutgers, The State University of New Jersey (US)
- Memorial Sloan Kettering Cancer Center (US)
- University of California, Davis (US)
- National Institutes of Health (US)
- Tri-Institutional PhD Program in Chemical Biology (US)
- Corresponding author
- Eastman, Peter
- Swails, Jason
- Chodera, John D.
- McGibbon, Robert T.
- Zhao, Yutong
- Beauchamp, Kyle A.
- Wang, Lee-Ping
- Simmonett, Andrew C.
- Harrigan, Matthew P.
- Stern, Chaya D.
- Wiewiora, Rafal P.
- Brooks, Bernard R.
- Pande, Vijay S.
- ORCID
- Venue
- PLOS Computational Biology
- Publisher
- Public Library of Science (PLoS)
- Published
- 2017-07-26
- Volume
- 13
- Issue
- 7
- Pages
- e1005659
- ISSN
- 1553-734X (ISSN-L)
- 1553-7358
- PMC
- PMC5549999
- OpenAlex
- W2949223833
- Cited by
- 2,748
- References
- 37
- Citations by year
- 2026: 380 · 2025: 455 · 2024: 472 · 2023: 432 · 2022: 330 · 2021: 301 · 2020: 197 · 2019: 105 · 2018: 65 · 2017: 11
- FWCI
- 119.89
- Type
- journal article
- Language
- en
- Update notices
- new_version (2017-08-09)
- Open access
- gold
- doi.org
- Licence
- public-domain
- creativecommons.org/publicdomain/zero/1.0/ (vor)
- Locations
- PLoS Computational Biology (publishedVersion, public-domain)
- bioRxiv (Cold Spring Harbor Laboratory) (acceptedVersion, cc-by)
- PubMed Central (publishedVersion, public-domain)
- PubMed (publishedVersion)
- RePEc: Research Papers in Economics (submittedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- eScholarship (California Digital Library) (submittedVersion)
- eScholarship (California Digital Library) (submittedVersion, public-domain)
- Figshare (submittedVersion, cc-by)
- Funders
- National Institutes of Health (US) (U54 GM072970, R01-GM062868)
- National Institutes of Health (P30 CA008748)
- Starr Foundation (US) (I8-A8-058)
- Topics
- Physical Sciences › Computer Science › Hardware and Architecture › Parallel Computing and Optimization Techniques
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Protein Structure and Dynamics
- Physical Sciences › Chemistry › Physical and Theoretical Chemistry › Advanced Physical and Chemical Molecular Interactions
- Keywords
- OpenMM, molecular dynamics simulation, extensibility, force field development, integration algorithms, simulation protocols, GPU acceleration, CPU simulation, high-performance algorithms
- MeSH terms
- Algorithms, Software, Computational Biology, Molecular Dynamics Simulation
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
Data sources
Bibliographic data: Semantic Scholar, OpenAlex, Crossref.
Contains information from the Semantic Scholar Academic Graph Datasets, made available under the ODC Attribution License (ODC-BY 1.0).
Semantic Scholar’s data is described in: Kinney, R., et al. (2023). The Semantic Scholar Open Data Platform. arXiv:2301.10140.
