

The DOI matches its Crossref record, with a small difference in title or year; 103 of the 104 references it cites were found in a registry or an index.
Martinize2 and Vermouth provide a unified framework for molecular topology generation
Open accessPublished version of a preprint
Abstract
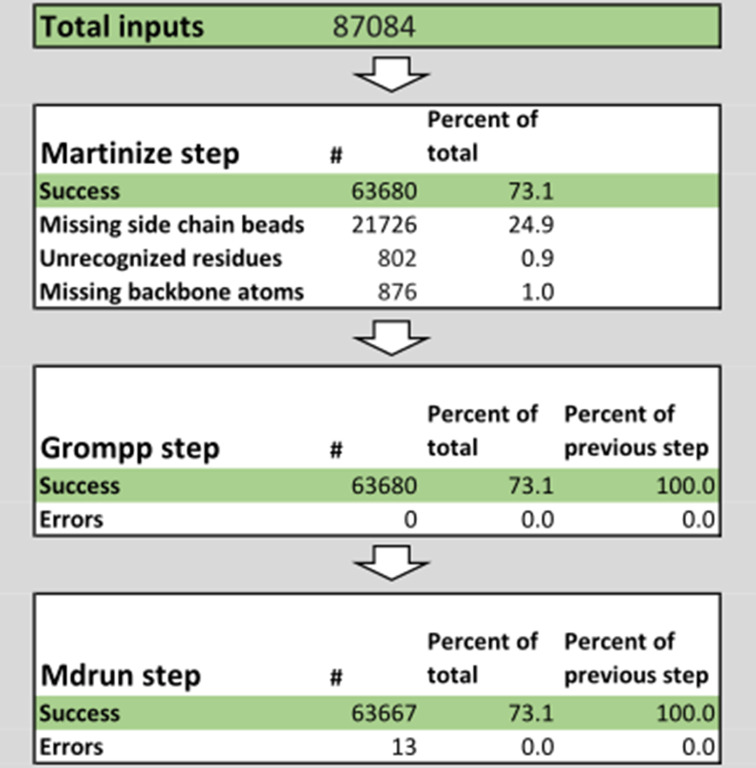
Ongoing advances in force field and computer hardware development enable the use of molecular dynamics (MD) to simulate increasingly complex systems with the ultimate goal of reaching cellular complexity. At the same time, rational design by high-throughput (HT) simulations is another forefront of MD. In these areas, the Martini coarse-grained force field, especially the latest version (i.e. v3), is being actively explored because it offers an enhanced spatial-temporal resolution. However, the automation tools for preparing simulations with the Martini force field, accompanying the previous version, were not designed for HT simulations or studies of complex cellular systems. Therefore, they become a major limiting factor. To address these shortcomings, we present the open-source Vermouth python library. Vermouth is designed to become the unified framework for developing programs, which prepare, run, and analyze Martini simulations of complex systems. To demonstrate the power of the Vermouth library, the Martinize2 program is showcased as a generalization of the martinize script, originally aimed to set up simulations of proteins. In contrast to the previous version, Martinize2 automatically handles protonation states in proteins and post-translation modifications, offers more options to fine-tune structural biases such as the elastic network (EN), and can convert non-protein molecules such as ligands. Finally, Martinize2 is used in two high-complexity benchmarks. The entire I-TASSER protein template database as well as a subset of 200,000 structures from the AlphaFold Protein Structure Database are converted to CG resolution and we illustrate how the checks on input structure quality can safeguard HT applications.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Highly accurate protein structure prediction with AlphaFold
Jumper, J., Evans, R., Pritzel, A., Green, T., Figurnov, M., Ronneberger, O., Tunyasuvunakool, K., Bates, R., Žídek, A., Potapenko, A., et al. (2021) · Nature · doi:10.1038/s41586-021-03819-2 · Cited by 43,764
SciPy 1.0: fundamental algorithms for scientific computing in Python
Virtanen, P., Gommers, R., Oliphant, T. E., Haberland, M., Reddy, T., Cournapeau, D., Burovski, E., Peterson, P., Weckesser, W., Bright, J., et al. (2020) · Nature Methods · doi:10.1038/s41592-019-0686-2 · Cited by 38,430
Molecular dynamics with coupling to an external bath
Berendsen, H. J. C., Postma, J. P. M., van Gunsteren, W. F., DiNola, A., & Haak, J. R. (1984) · The Journal of Chemical Physics · doi:10.1063/1.448118 · Cited by 31,057
Abraham, M. J., Murtola, T., Schulz, R., Páll, S., Smith, J. C., Hess, B., & Lindahl, E. (2015) · SoftwareX · doi:10.1016/j.softx.2015.06.001 · Cited by 27,190
Harris, C. R., Millman, K. J., van der Walt, S. J., Gommers, R., Virtanen, P., Cournapeau, D., Wieser, E., Taylor, J., Berg, S., Smith, N. J., et al. (2020) · Nature · doi:10.1038/s41586-020-2649-2 · Cited by 23,826 · cited 1×
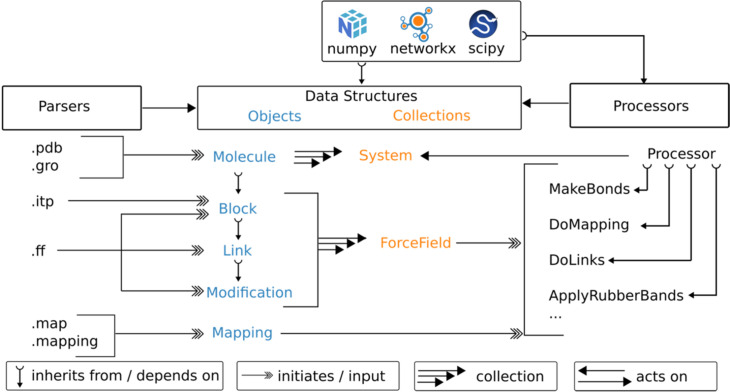
“In addition, the library relies on only three permissibly licensed open-software projects, namely numpy ( Harris et al., 2020 ), scipy ( Virtanen et al., 2020 ), and networkx ( Hagberg et al., 2008 ).”
Polymorphic transitions in single crystals: A new molecular dynamics method
Parrinello, M., & Rahman, A. (1981) · Journal of Applied Physics · doi:10.1063/1.328693 · Cited by 20,009
The FAIR Guiding Principles for scientific data management and stewardship
Wilkinson, M. D., Dumontier, M., Aalbersberg, I. J., Appleton, G., Axton, M., Baak, A., Blomberg, N., Boiten, J.-W., da Silva Santos, L. B., Bourne, P. E., et al. (2016) · Scientific Data · doi:10.1038/sdata.2016.18 · Cited by 18,423 · cited 2×
“( Chue Hong et al., 2022 ; Wilkinson et al., 2016 ; Alibay et al., 2022 ) The FAIR principles Wilkinson et al., 2016 , originally designed to improve data management and reproducibility in science, have recently been extended to research software in a more general sense.”
Canonical sampling through velocity rescaling
Bussi, G., Donadio, D., & Parrinello, M. (2007) · The Journal of Chemical Physics · doi:10.1063/1.2408420 · Cited by 18,327
Scalable molecular dynamics with NAMD
Phillips, J. C., Braun, R., Wang, W., Gumbart, J., Tajkhorshid, E., Villa, E., Chipot, C., Skeel, R. D., Kalé, L., & Schulten, K. (2005) · Journal of Computational Chemistry · doi:10.1002/jcc.20289 · Cited by 16,914
Kabsch, W., & Sander, C. (1983) · Biopolymers · doi:10.1002/bip.360221211 · Cited by 15,320
CHARMM: A program for macromolecular energy, minimization, and dynamics calculations
Brooks, B., Bruccoleri, R., Olafson, B. D., States, D., Swaminathan, S., & Karplus, M. (1983) · Journal of Computational Chemistry · doi:10.1002/jcc.540040211 · Cited by 13,736
Reducibility among Combinatorial Problems
Karp, R. M. (1972) · Complexity of Computer Computations · doi:10.1007/978-1-4684-2001-2_9 · Cited by 11,511
The Amber biomolecular simulation programs
Case, D. A., Cheatham, T. E., Darden, T., Gohlke, H., Luo, R., Merz, K. M., Onufriev, A., Simmerling, C., Wang, B., & Woods, R. J. (2005) · Journal of Computational Chemistry · doi:10.1002/jcc.20290 · Cited by 9,535
CHARMM‐GUI: A web‐based graphical user interface for CHARMM
Jo, S., Kim, T., Iyer, V. G., & Im, W. (2008) · Journal of Computational Chemistry · doi:10.1002/jcc.20945 · Cited by 9,284
CHARMM: The biomolecular simulation program
Brooks, B. R., Brooks, C. L., MacKerell, A. D., Nilsson, L., Petrella, R. J., Roux, B., Won, Y., Archontis, G., Bartels, C., Boresch, S., et al. (2009) · Journal of Computational Chemistry · doi:10.1002/jcc.21287 · Cited by 9,115
Varadi, M., Anyango, S., Deshpande, M., Nair, S., Natassia, C., Yordanova, G., Yuan, D., Stroe, O., Wood, G., Laydon, A., et al. (2022) · Nucleic Acids Research · doi:10.1093/nar/gkab1061 · Cited by 7,850
Exploring Network Structure, Dynamics, and Function Using NetworkX
Hagberg, A. A., Schult, D. A., & Swart, P. J. (2008) · Proceedings of the Python in Science Conference · doi:10.25080/tcwv9851 · Cited by 7,477
CHARMM36m: an improved force field for folded and intrinsically disordered proteins
Huang, J., Rauscher, S., Nawrocki, G., Ran, T., Feig, M., de Groot, B. L., Grubmüller, H., & MacKerell, A. D. (2016) · Nature Methods · doi:10.1038/nmeth.4067 · Cited by 7,334
The complexity of theorem-proving procedures
Cook, S. A. (1971) · Proceedings of the third annual ACM symposium on Theory of computing - STOC '71 · doi:10.1145/800157.805047 · Cited by 6,385
Improved side-chain torsion potentials for the Amber ff99SB protein force field
Lindorff-Larsen, K., Piana, S., Palmo, K., Maragakis, P., Klepeis, J., Dror, R., & Shaw, D. (2010) · Proteins: Structure, Function, and Bioinformatics · doi:10.1002/prot.22711 · Cited by 6,056
The MARTINI Force Field: Coarse Grained Model for Biomolecular Simulations
Marrink, . J., Risselada, H. J., Yefimov, S., Tieleman, D. P., & de Vries, A. H. (2007) · The Journal of Physical Chemistry B · doi:10.1021/jp071097f · Cited by 5,641
The I-TASSER Suite: protein structure and function prediction
Yang, J., Yan, R., Roy, A., Xu, D., Poisson, J., & Zhang, Y. (2015) · Nature Methods · doi:10.1038/nmeth.3213 · Cited by 5,596
PROPKA3: Consistent Treatment of Internal and Surface Residues in Empirical p K a Predictions
Olsson, M. H. M., Søndergaard, C. R., Rostkowski, M., & Jensen, J. H. (2011) · Journal of Chemical Theory and Computation · doi:10.1021/ct100578z · Cited by 4,425
Lee, J.-M., Cheng, X., Swails, J. M., Yeom, M. S., Eastman, P., Lemkul, J. A., Wei, S., Buckner, J., Jeong, J. C., Qi, Y., et al. (2015) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.5b00935 · Cited by 4,252
OpenMM 7: Rapid development of high performance algorithms for molecular dynamics
Eastman, P., Swails, J., Chodera, J. D., McGibbon, R. T., Zhao, Y., Beauchamp, K. A., Wang, L.-P., Simmonett, A. C., Harrigan, M. P., Stern, C. D., et al. (2017) · PLOS Computational Biology · doi:10.1371/journal.pcbi.1005659 · Cited by 2,748
The MARTINI Coarse-Grained Force Field: Extension to Proteins
Monticelli, L., Kandasamy, S. K., Périole, X., Larson, R. G., Tieleman, D. P., & Marrink, . J. (2008) · Journal of Chemical Theory and Computation · doi:10.1021/ct700324x · Cited by 2,521
Automation of the CHARMM General Force Field (CGenFF) I: bond perception and atom typing
Vanommeslaeghe, K., & MacKerell, A. D. (2012) · Journal of Chemical Information and Modeling · doi:10.1021/ci300363c · Cited by 2,035
An Automated Force Field Topology Builder (ATB) and Repository: Version 1.0
Malde, A. K., Zuo, L., Breeze, M., Stroet, M., Poger, D., Nair, P. C., Oostenbrink, C., & Mark, A. E. (2011) · Journal of Chemical Theory and Computation · doi:10.1021/ct200196m · Cited by 1,897
Author Correction: SciPy 1.0: fundamental algorithms for scientific computing in Python
Contributors, S. 1. 0., Virtanen, P., Gommers, R., Oliphant, T. E., Haberland, M., Reddy, T. J., Cournapeau, D., Burovski, E., Peterson, P., Weckesser, W., et al. (2020) · Nature Methods · doi:10.1038/s41592-020-0772-5 · Cited by 1,848
Anandakrishnan, R., Aguilar, B. H., & Onufriev, A. (2012) · Nucleic Acids Res. · doi:10.1093/nar/gks375 · Cited by 1,736
Jorgensen, W. L., & Tirado‐Rives, J. (2005) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.0408037102 · Cited by 1,666
LigParGen web server: an automatic OPLS-AA parameter generator for organic ligands
Dodda, L. S., de Vaca, I. C., Tirado-Rives, J., & Jorgensen, W. L. (2017) · Nucleic Acids Res. · doi:10.1093/nar/gkx312 · Cited by 1,600
Martini 3: A general purpose force field for coarse-grained molecular dynamics
Souza, P. C. T., Alessandri, R., Barnoud, J., Thallmair, S., Faustino, I., Grünewald, F., Patmanidis, I., Abdizadeh, H., Bruininks, B. M. H., Wassenaar, T. A., et al. (2021) · Nature Methods · doi:10.1038/s41592-021-01098-3 · Cited by 1,551
Self-citation‘Improved parameters for the Martini coarse-grained protein force field’
de Jong, D. H., Singh, G., Bennett, W. F. D., Arnarez, C., Wassenaar, T. A., Schäfer, L. V., Periole, X., Tieleman, D. P., & Marrink, S. J. (2013) · Journal of Chemical Theory and Computation · doi:10.1021/ct300646g · Cited by 1,486
Self-citationA (sub)graph isomorphism algorithm for matching large graphs
Cordella, L. P., Foggia, P., Sansone, C., & Vento, M. (2004) · IEEE Transactions on Pattern Analysis and Machine Intelligence · doi:10.1109/tpami.2004.75 · Cited by 1,486
Bjelkmar, P., Larsson, P., Cuendet, M. A., Hess, B., & Lindahl, E. R. (2010) · Journal of Chemical Theory and Computation · doi:10.1021/ct900549r · Cited by 1,308
Tackling Exascale Software Challenges in Molecular Dynamics Simulations with GROMACS
Páll, S., Abraham, M., Kutzner, C., Hess, B., & Lindahl, E. R. (2015) · Lecture notes in computer science · doi:10.1007/978-3-319-15976-8_1 · Cited by 1,215
Coarse-Grained Protein Models and Their Applications
Kmiecik, S., Gront, D., Koliński, M., Wieteska, Ł., Badaczewska-Dawid, A. E., & Koliński, A. (2016) · Chemical Reviews · doi:10.1021/acs.chemrev.6b00163 · Cited by 1,113
pKa's of ionizable groups in proteins: atomic detail from a continuum electrostatic model
Bashford, D., & Karplus, M. (1990) · Biochemistry · doi:10.1021/bi00496a010 · Cited by 1,089
A series of PDB-related databanks for everyday needs
Touw, W. G., Baakman, C. A. B., Black, J., Beek, T. A. H. T., Krieger, E., Joosten, R. P., & Vriend, G. (2014) · Nucleic Acids Research · doi:10.1093/nar/gku1028 · Cited by 941
1.14*CM1A-LBCC: Localized Bond-Charge Corrected CM1A Charges for Condensed-Phase Simulations
Dodda, L. S., Vilseck, J. Z., Tirado‐Rives, J., & Jorgensen, W. L. (2017) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.7b00272 · Cited by 885
Wassenaar, T. A., Pluháčková, K., Böckmann, R. A., Marrink, . J., & Tieleman, D. P. (2013) · Journal of Chemical Theory and Computation · doi:10.1021/ct400617g · Cited by 727
Self-citationComputational Modeling of Realistic Cell Membranes
Marrink, . J., Corradi, V., Souza, P. C. T., Ingólfsson, H. I., Tieleman, D. P., & Sansom, M. S. P. (2019) · Chemical Reviews · doi:10.1021/acs.chemrev.8b00460 · Cited by 715
Self-citationPériole, X., Cavalli, M., Marrink, . J., & Ceruso, M. A. (2009) · Journal of Chemical Theory and Computation · doi:10.1021/ct9002114 · Cited by 698
STUDIES ON PROTEIN FOLDING, UNFOLDING AND FLUCTUATIONS BY COMPUTER SIMULATION
Taketomi, H., Ueda, Y., & Gō, N. (1975) · International journal of peptide & protein research · doi:10.1111/j.1399-3011.1975.tb02465.x · Cited by 679
CHARMM-GUI Martini Maker for Coarse-Grained Simulations with the Martini Force Field
Qi, Y., Ingólfsson, H. I., Cheng, X., Lee, J., Marrink, S. J., & Im, W. (2015) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.5b00513 · Cited by 494
Martini straight: Boosting performance using a shorter cutoff and GPUs
de Jong, D. H., Baoukina, S., Ingólfsson, H. I., & Marrink, . J. (2015) · Computer Physics Communications · doi:10.1016/j.cpc.2015.09.014 · Cited by 437
CHARMM‐GUI 10 years for biomolecular modeling and simulation
Jo, S., Cheng, X., Lee, J., Kim, S., Park, S., Patel, D. S., Beaven, A. H., Lee, K. I., Rui, H., Park, S., et al. (2016) · Journal of Computational Chemistry · doi:10.1002/jcc.24660 · Cited by 381
An Improved Algorithm for Matching Large Graphs
Cordella, L. P., Foggia, P., Sansone, C., & Vento, M. (2001) · Cited by 358
Martini Coarse-Grained Force Field: Extension to DNA
Uusitalo, J. J., Ingólfsson, H. I., Akhshi, P., Tieleman, D. P., & Marrink, . J. (2015) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.5b00286 · Cited by 330
Martini coarse-grained force field: Extension to RNA
Uusitalo, J. J., Ingólfsson, H. I., Marrink, S. J., & Faustino, I. (2017) · Biophysical Journal · doi:10.1016/j.bpj.2017.05.043 · Cited by 259 · cited 2×
“…et al., 2017 ; Girard et al., 2019 ; Jo et al., 2017 ; Qi et al., 2015 ; Malde et al., 2011 ; Canzar et al., 2013 ; Jorgensen and Tirado-Rives, 2005 ; Dodda et al., 2017b ; Dodda et al., 2017a ; Vanommeslaeghe and MacKerell, 2012 ; Uusitalo et al., 2015 ; Jo et al., 2008 ; Uusitalo et al., 2017 ).”
Two decades of Martini: Better beads, broader scope
Marrink, S. J., Monticelli, L., Melo, M. N., Alessandri, R., Tieleman, D. P., & Souza, P. C. T. (2023) · WIREs Computational Molecular Science · doi:10.1002/wcms.1620 · Cited by 255
Self-citationProtein–ligand binding with the coarse-grained Martini model
Souza, P. C. T., Thallmair, S., Conflitti, P., Ramírez-Palacios, C., Alessandri, R., Raniolo, S., Limongelli, V., & Marrink, . J. (2020) · Nature Communications · doi:10.1038/s41467-020-17437-5 · Cited by 246
Self-citationEnumerating all connected maximal common subgraphs in two graphs
Koch, I. (2001) · Theoretical Computer Science · doi:10.1016/s0304-3975(00)00286-3 · Cited by 245
A subgraph isomorphism algorithm and its application to biochemical data
Bonnici, V., Giugno, R., Pulvirenti, A., Shasha, D. E., & Ferro, A. (2013) · BMC Bioinformatics · doi:10.1186/1471-2105-14-s7-s13 · Cited by 220
Poma, A. B., Cieplak, M., & Theodorakis, P. E. (2017) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.6b00986 · Cited by 219
Martini 3 Coarse‐Grained Force Field: Small Molecules
Alessandri, R., Barnoud, J., Gertsen, A. S., Patmanidis, I., de Vries, A. H., Souza, P. C. T., & Marrink, . J. (2021) · Advanced Theory and Simulations · doi:10.1002/adts.202100391 · Cited by 213
Self-citationA maximum common substructure-based algorithm for searching and predicting drug-like compounds
Cao, Y., Jiang, T., & Girke, T. (2008) · Bioinformatics · doi:10.1093/bioinformatics/btn186 · Cited by 211
Constant pH Molecular Dynamics in Explicit Solvent with λ-Dynamics
Donnini, S., Tegeler, F., Groenhof, G., & Grubmüller, H. (2011) · Journal of Chemical Theory and Computation · doi:10.1021/ct200061r · Cited by 207
High-Throughput All-Atom Molecular Dynamics Simulations Using Distributed Computing
Buch, I., Harvey, M., Giorgino, T., Anderson, D. P., & de Fabritiis, G. (2010) · Journal of Chemical Information and Modeling · doi:10.1021/ci900455r · Cited by 183
Polyply; a python suite for facilitating simulations of macromolecules and nanomaterials
Grünewald, F., Alessandri, R., Kroon, P. C., Monticelli, L., Souza, P. C. T., & Marrink, . J. (2022) · Nature Communications · doi:10.1038/s41467-021-27627-4 · Cited by 178
Self-citationA multiscale coarse-grained model of the SARS-CoV-2 virion
Yu, A., Pak, A. J., He, P., Monje‐Galvan, V., Casalino, L., Gaieb, Z., Dommer, A. C., Amaro, R. E., & Voth, G. A. (2020) · Biophysical Journal · doi:10.1016/j.bpj.2020.10.048 · Cited by 169
Charge Group Partitioning in Biomolecular Simulation
Canzar, S., El-Kebir, M., Pool, R., Elbassioni, K., Malde, A. K., Mark, A. E., Geerke, D. P., Stougie, L., & Klau, G. W. (2013) · Journal of Computational Biology · doi:10.1089/cmb.2012.0239 · Cited by 161
Transferable MARTINI Model of Poly(ethylene Oxide)
Grünewald, F., Rossi, G., de Vries, A. H., Marrink, . J., & Monticelli, L. (2018) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.8b04760 · Cited by 157
Self-citationMolecular dynamics simulation of an entire cell
Stevens, J. A., Grünewald, F., van Tilburg, P. A. M., König, M., Gilbert, B. R., Brier, T. A., Thornburg, Z. R., Luthey-Schulten, Z., & Marrink, S. J. (2023) · Frontiers in Chemistry · doi:10.3389/fchem.2023.1106495 · Cited by 156
Self-citationBackmapping triangulated surfaces to coarse-grained membrane models
Pezeshkian, W., König, M., Wassenaar, T. A., & Marrink, . J. (2020) · Nature Communications · doi:10.1038/s41467-020-16094-y · Cited by 156
Self-citationScalable Constant pH Molecular Dynamics in GROMACS
Aho, N., Buslaev, P., Jansen, A., Bauer, P., Groenhof, G., & Hess, B. (2022) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.2c00516 · Cited by 155
GROMACS in the Cloud: A Global Supercomputer to Speed Up Alchemical Drug Design
Kutzner, C., Kniep, C., Cherian, A., Nordstrom, L., Grubmüller, H., de Groot, B. L., & Gapsys, V. (2022) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.2c00044 · Cited by 142
All-atom Continuous Constant pH Molecular Dynamics With Particle Mesh Ewald and Titratable Water
Huang, Y.-D., Chen, W., Wallace, J. A., & Shen, J. (2016) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.6b00552 · Cited by 132
MARTINI Coarse-Grained Models of Polyethylene and Polypropylene
Panizon, E., Bochicchio, D., Monticelli, L., & Rossi, G. (2015) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.5b03611 · Cited by 124
Common subgraph isomorphism detection by backtracking search
Krissinel, E., & Henrick, K. (2004) · Software Practice and Experience · doi:10.1002/spe.588 · Cited by 110
Dommer, A. C., Casalino, L., Kearns, F. L., Rosenfeld, M. A., Wauer, N. A., Ahn, S.-H., Russo, J. D., Oliveira, A. S. F., Morris, C. K., Bogetti, A. T., et al. (2022) · The International Journal of High Performance Computing Applications · doi:10.1177/10943420221128233 · Cited by 110
SIRAH tools: mapping, backmapping and visualization of coarse-grained models
Machado, M. R., & Pantano, S. (2016) · Bioinform. · doi:10.1093/bioinformatics/btw020 · Cited by 107
Danne, R., Poojari, C., Martinez‐Seara, H., Rissanen, S., Lolicato, F., Róg, T., & Vattulainen, I. (2017) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.7b00237 · Cited by 96
Graph-Based Approach to Systematic Molecular Coarse-Graining
Webb, M. A., Delannoy, J.-Y. P., & Pablo, J. (2018) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.8b00920 · Cited by 93
Titratable Martini model for constant pH simulations
Grünewald, F., Souza, P. C. T., Abdizadeh, H., Barnoud, J., de Vries, A. H., & Marrink, . J. (2020) · The Journal of Chemical Physics · doi:10.1063/5.0014258 · Cited by 93
Self-citationImproved Side Chain Dynamics in MARTINI Simulations of Protein–Lipid Interfaces
Herzog, F. A., Braun, L., Schoen, I., & Viola Vogel, B. (2016) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.6b00122 · Cited by 86
Empereur‐mot, C., Pesce, L., Doni, G., Bochicchio, D., Capelli, R., Perego, C., & Pavan, G. M. (2020) · ACS Omega · doi:10.1021/acsomega.0c05469 · Cited by 82
Constant pH simulations with the coarse-grained MARTINI model — Application to oleic acid aggregates
Bennett, W. F., Chen, A. W., Donnini, S., Groenhof, G., & Tieleman, D. P. (2013) · Canadian Journal of Chemistry · doi:10.1139/cjc-2013-0010 · Cited by 81
FAIR Principles for Research Software (FAIR4RS Principles)
Chue Hong, N. P., Katz, D. S., Barker, M., Lamprecht, A.-L., Martínez-Ortiz, C., Psomopoulos, F. E., Harrow, J., Castro, L. J., Gruenpeter, M., Martinez, P. A., et al. (2021) · Edinburgh Research Explorer (University of Edinburgh) · doi:10.15497/rda00068 · Cited by 81
Whole-Cell Models and Simulations in Molecular Detail
Feig, M., & Sugita, Y. (2019) · Annual Review of Cell and Developmental Biology · doi:10.1146/annurev-cellbio-100617-062542 · Cited by 78
Nnyigide, O. S., Nnyigide, T. O., Lee, S., & Hyun, K. (2022) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.2c00571 · Cited by 76
Challenges in structural approaches to cell modeling
Im, W., Liang, J., Olson, A. J., Zhou, H., Vajda, S., & Vakser, I. A. (2016) · Journal of Molecular Biology · doi:10.1016/j.jmb.2016.05.024 · Cited by 70
Molecular architecture and dynamics of SARS-CoV-2 envelope by integrative modeling
Pezeshkian, W., Grünewald, F., Narykov, O., Lu, S., Arkhipova, V., Solodovnikov, A., Wassenaar, T. A., Marrink, S. J., & Korkin, D. (2023) · Structure · doi:10.1016/j.str.2023.02.006 · Cited by 60
Self-citationEncoding and selecting coarse-grain mapping operators with hierarchical graphs
Chakraborty, M., Xu, C., & White, A. D. (2018) · The Journal of Chemical Physics · doi:10.1063/1.5040114 · Cited by 54
Perspectives on High-Throughput Ligand/Protein Docking With Martini MD Simulations
Souza, P. C. T., Limongelli, V., Wu, S., Marrink, . J., & Monticelli, L. (2021) · Frontiers in Molecular Biosciences · doi:10.3389/fmolb.2021.657222 · Cited by 52
Self-citationFacilitating CG Simulations with MAD: The MArtini Database Server
Hilpert, C., Beranger, L., Souza, P. C. T., Vainikka, P. A., Nieto, V., Marrink, S. J., Monticelli, L., & Launay, G. (2023) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.2c01375 · Cited by 51
Self-citationHoubraken, M., Demeyer, S., Michoel, T., Audenaert, P., Colle, D., & Pickavet, M. (2014) · PLoS ONE · doi:10.1371/journal.pone.0097896 · Cited by 47
Pedersen, K. B., Borges-Araújo, L., Stange, A. D., Souza, P. C. T., Marrink, . J., & Schiøtt, B. (2024) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.4c00553 · Cited by 41
Self-citationHoobas: A highly object-oriented builder for molecular dynamics
Girard, M., Ehlen, A., Shakya, A., Bereau, T., & de la Cruz, M. O. (2019) · Computational Materials Science · doi:10.1016/j.commatsci.2019.05.003 · Cited by 37
Demeyer, S., Michoel, T., Fostier, J., Audenaert, P., Pickavet, M., & Demeester, P. M. A. (2013) · PLoS ONE · doi:10.1371/journal.pone.0061183 · Cited by 34
Sousa, F. M., Lima, L. M. P., Arnarez, C., Pereira, M. M., & Melo, M. N. (2021) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.0c01077 · Cited by 30
SIDUS—the solution for extreme deduplication of an operating system
QuemenerEmmanuel, & CorvellecMarianne (2013) · Linux journal · doi:10.5555/2555789.2555792 · Cited by 30
Integrative structural modelling and visualisation of a cellular organelle
Autin, L., Barbaro, B. A., Jewett, A. I., Ekman, A. A., Verma, S., Olson, A. J., & Goodsell, D. S. (2022) · QRB Discovery · doi:10.1017/qrd.2022.10 · Cited by 28
When computational pipelines go ‘clank’
Marx, V. (2020) · Nature Methods · doi:10.1038/s41592-020-0886-9 · Cited by 19
MartiniGlass: a Tool for Enabling Visualization of Coarse-Grained Martini Topologies
Brasnett, C., & Marrink, . J. (2025) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.4c02277 · Cited by 17
Self-citationMartini 3 OliGomers: A Scalable Approach for Multimers and Fibrils in GROMACS
Korshunova, K., Kiuru, J., Liekkinen, J., Enkavi, G., Vattulainen, I., & Bruininks, B. M. H. (2024) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.4c00677 · Cited by 14
SIDUS—the solution for extreme deduplication of an operating system
Quémener, E., & Corvellec, M. (2013) · Linux journal · Cited by 14
CGsmiles: A Versatile Line Notation for Molecular Representations across Multiple Resolutions
Grünewald, F., Seute, L., Alessandri, R., König, M., & Kroon, P. C. (2025) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.5c00064 · Cited by 13
Self-citationEnumerating common molecular substructuresPreprint
Engler, M. S., El-Kebir, M., Mulder, J., Mark, A. E., Geerke, D. P., & Klau, G. W. (2017) · doi:10.7287/peerj.preprints.3250v1 · Cited by 11
Multiple-Choice Knapsack for Assigning Partial Atomic Charges in Drug-Like Molecules
Engler, M. S., Caron, B., Veen, L. E., Geerke, D. P., Mark, A. E., & Klau, G. W. (2018) · Centrum Wiskunde & Informatica (CWI), the national research institute for mathematics and computer science in the Netherlands · doi:10.4230/lipics.wabi.2018.16 · Cited by 11
BioExcel Whitepaper on Scientific Software Development
Abraham, M., Melquiond, A. S. J., Ippoliti, E., Gapsys, V., Hess, B., Trellet, M. E., Rodrigues, J., Laure, E., Apostolov, R., de Groot, B. L., et al. (2018) · Zenodo (CERN European Organization for Nuclear Research) · doi:10.5281/zenodo.1194633 · Cited by 6
Alibay, I., Barnoud, J., Beckstein, O., Gowers, R. J., Naughton, F. B., & Wang, L. (2022) · Figshare · doi:10.6084/m9.figshare.20520726 · Cited by 3
Self-citation
Research works printed but not linked: 3
- Chung, 2010, Graph theory in the information age, Notices of the American Mathematical Society, 57, 726
- Kroon PC, Barnoud J, Brasnett C, Grunewald F, Peter M, Tadema M, Liu X, Araújo LB, marrink-lab, Tsjerk, Romumrn. 2025. Vermouth-martinize. Software Heritage. swh:1:rev:a3037bc2544daa0faa8ba105b9670a2659885d43. https://archive.softwareheritage.org/swh:1:dir:270aae9780a9b8f07c0c1411175f4e982bc0b539;origin=https://github.com/marrink-lab/vermouth-martinize;visit=swh:1:snp:155374469e37d66bb87fc5832a0e0f314e27ca85;anchor=swh:1:rev:a3037bc2544daa0faa8ba105b9670a2659885d43.
- marrink-lab. 2025. Martinize-examples. Software Heritage. swh:1:rev:78a81b00ff8bc9798386c442251a14f568e8f8eb. https://archive.softwareheritage.org/swh:1:dir:0869dc8a9197f72f91847dbb5739f97fb20576bd;origin=https://github.com/marrink-lab/martinize-examples;visit=swh:1:snp:85479e72dfea864eca7b37f1eb7390153820e6eb;anchor=swh:1:rev:78a81b00ff8bc9798386c442251a14f568e8f8eb.
Cited by
Sorted by citations
Martini 3 Coarse-Grained Force Field for Cholesterol
Borges-Araújo, L., Borges-Araújo, A. C., Öztürk, T. N., Ramirez-Echemendía, D. P., Fábián, B., Carpenter, T. S., Thallmair, S., Barnoud, J., Ingólfsson, H. I., Hummer, G., et al. (2023) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.3c00547 · Cited by 87
Self-citationMartini3-IDP: improved Martini 3 force field for disordered proteins
Wang, L., Brasnett, C., Borges-Araújo, L., Souza, P. C. T., & Marrink, . J. (2025) · Nature Communications · doi:10.1038/s41467-025-58199-2 · Cited by 70
Self-citationFacilitating CG Simulations with MAD: The MArtini Database Server
Hilpert, C., Beranger, L., Souza, P. C. T., Vainikka, P. A., Nieto, V., Marrink, S. J., Monticelli, L., & Launay, G. (2023) · Journal of Chemical Information and Modeling · doi:10.1021/acs.jcim.2c01375 · Cited by 51
Self-citationAccurate model and ensemble refinement using cryo-electron microscopy maps and Bayesian inference
Hoff, S., Thomasen, F. E., Lindorff‐Larsen, K., & Bonomi, M. (2024) · PLoS Computational Biology · doi:10.1371/journal.pcbi.1012180 · Cited by 43
Hexokinase-I directly binds to a charged membrane-buried glutamate of mitochondrial VDAC1 and VDAC2
Bieker, S., Timme, M., Woge, N., Hassan, D., Brown, C. M., Marrink, . J., Melo, M. N., & Holthuis, J. C. M. (2025) · Communications Biology · doi:10.1038/s42003-025-07551-9 · Cited by 30
Metadata
- Authors
- Kroon, P. C., Grünewald, F., Barnoud, J., van Tilburg, M., Brasnett, C., Souza, P. C., Wassenaar, T. A., & Marrink, S. J.
- Affiliations
- University of Groningen (NL)
- École Normale Supérieure de Lyon (FR)
- Corresponding author
- Grünewald, Fabian
- Marrink, Siewert J
- Venue
- eLife
- Publisher
- eLife Sciences Publications, Ltd
- Published
- 2025-11-20
- Volume
- 12
- ISSN
- 2050-084X (ISSN-L)
- PMC
- PMC12634043
- OpenAlex
- W7106126080
- Cited by
- 124
- References
- 104
- Citations by year
- 2026: 73 · 2025: 22
- Type
- posted content
- Language
- en
- Open access
- gold
- doi.org
- Licence
- Locations
- eLife (publishedVersion, cc-by)
- Europe PMC (PubMed Central) (publishedVersion, cc-by)
- PubMed (publishedVersion)
- University of Groningen research database (University of Groningen / Centre for Information Technology) (publishedVersion, cc-by)
- arXiv (Cornell University) (submittedVersion)
- HAL (submittedVersion, cc-by)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion, cc-by-sa)
- arXiv (Cornell University) (cc-by)
- Funders
- European Research Council (COMP-MICR-CROW-MEM)
- Novo Nordisk (BOUNDLESS)
- Klaus Tschira Foundation (Independent Postdoc)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Protein Structure and Dynamics
- Physical Sciences › Chemistry › Physical and Theoretical Chemistry › Advanced Physical and Chemical Molecular Interactions
- Life Sciences › Pharmacology, Toxicology and Pharmaceutics › Drug Discovery › Computational Drug Discovery Methods
- Keywords
- Martini coarse-grained force field, vermouth, high-throughput molecular dynamics, protonation states, post-translational modification, elastic networks, AlphaFold structures
- MeSH terms
- Proteins, Software, Molecular Dynamics Simulation
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
