

The DOI matches its Crossref record; 73 of the 73 references it cites were found in a registry or an index.
Feature selection and dimension reduction for single-cell RNA-Seq based on a multinomial model
Open accessPublished version of a preprint
Abstract
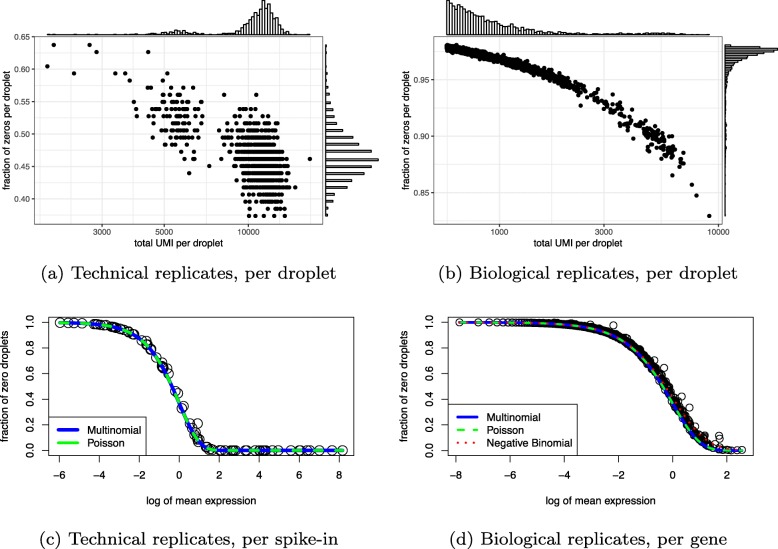
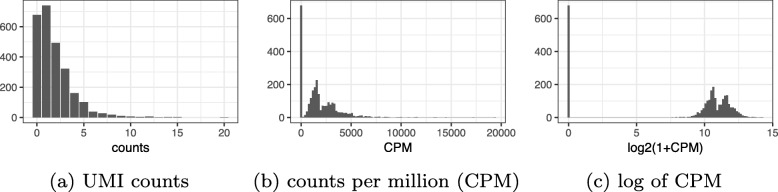
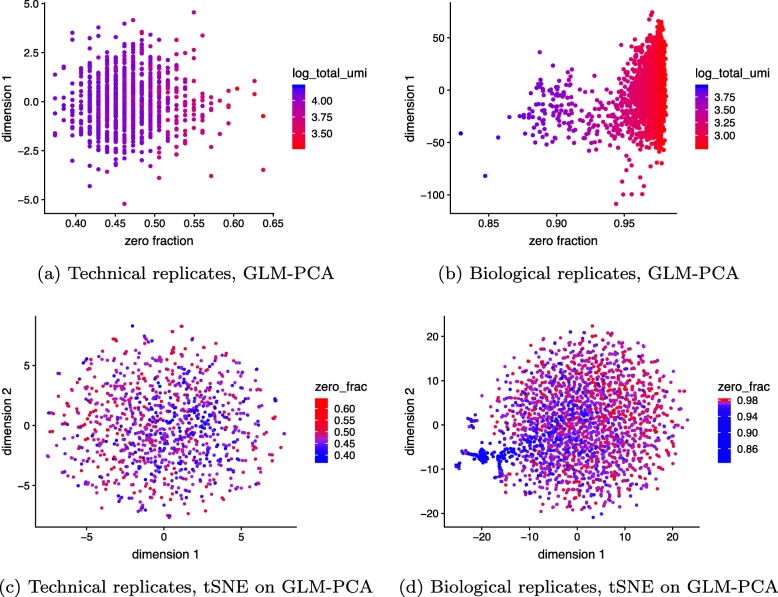
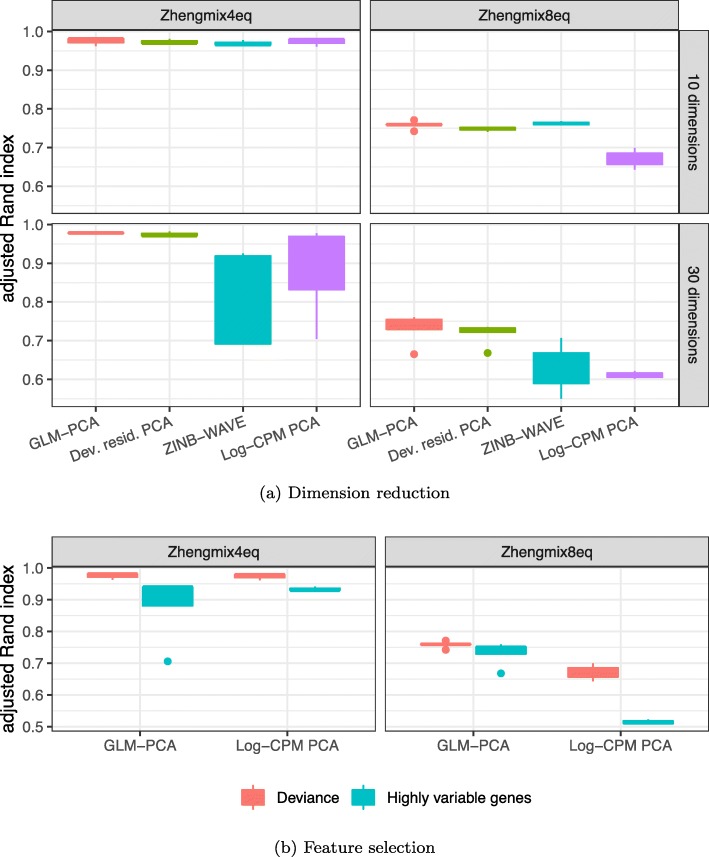
Single-cell RNA-Seq (scRNA-Seq) profiles gene expression of individual cells. Recent scRNA-Seq datasets have incorporated unique molecular identifiers (UMIs). Using negative controls, we show UMI counts follow multinomial sampling with no zero inflation. Current normalization procedures such as log of counts per million and feature selection by highly variable genes produce false variability in dimension reduction. We propose simple multinomial methods, including generalized principal component analysis (GLM-PCA) for non-normal distributions, and feature selection using deviance. These methods outperform the current practice in a downstream clustering assessment using ground truth datasets.
Overview
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
The claims and statistics of this paper haven’t been checked yet.
Run the check — free, no account needed. It reads the open-access text and checks the sources it cites.
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
In the citing paper
“Deviance identifies highly informative genes by fitting a gene-wise model that assumes constant expression across all cells and quantifying which genes violate this assumption47.”
From Best practices for single-cell analysis across modalities (2023) · cites this paper as [47]
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
The self-check needs the paper’s full text.
Citations
Cites
Algorithm AS 136: A K-Means Clustering Algorithm
Hartigan, J. A., & Wong, M. A. (1979) · Applied Statistics · doi:10.2307/2346830 · Cited by 14,523
Integrating single-cell transcriptomic data across different conditions, technologies, and species
Butler, A., Hoffman, P., Smibert, P., Papalexi, E., & Satija, R. (2018) · Nature Biotechnology · doi:10.1038/nbt.4096 · Cited by 13,174
Analysis of a complex of statistical variables into principal components.
Hotelling, H. (1933) · Journal of Educational Psychology · doi:10.1037/h0071325 · Cited by 9,691
Advances in Neural Information Processing Systems 14
(2002) · The MIT Press eBooks · doi:10.7551/mitpress/1120.001.0001 · Cited by 9,187
Hubert, L., & Arabie, P. (1985) · Journal of Classification · doi:10.1007/bf01908075 · Cited by 8,219
Massively parallel digital transcriptional profiling of single cells
Zheng, G. X. Y., Terry, J. M., Belgrader, P., Ryvkin, P., Bent, Z. W., Wilson, R., Ziraldo, S. B., Wheeler, T. D., McDermott, G. P., Zhu, J., et al. (2017) · Nature Communications · doi:10.1038/ncomms14049 · Cited by 7,353
Trapnell, C., Cacchiarelli, D., Grimsby, J., Pokharel, P., Li, S., Morse, M., Lennon, N. J., Livak, K. J., Mikkelsen, T. S., & Rinn, J. L. (2014) · Nature Biotechnology · doi:10.1038/nbt.2859 · Cited by 7,314
Highly parallel genome-wide expression profiling of individual cells using nanoliter droplets.
Macosko, E. Z., Basu, A., Satija, R., Nemesh, J., Shekhar, K., Goldman, M., Tirosh, I., Bialas, A. R., Kamitaki, N., Martersteck, E. M., et al. (2015) · Cell · doi:10.1016/j.cell.2015.05.002 · Cited by 7,110
UMAP: Uniform Manifold Approximation and Projection for Dimension Reduction
McInnes, L., Healy, J., & Melville, J. (2018) · arXiv · doi:10.48550/arxiv.1802.03426 · Cited by 7,101
Hafemeister, C., & Satija, R. (2019) · Genome Biology · doi:10.1186/s13059-019-1874-1 · Cited by 4,479 · cited 1×
“Recently, Pearson residuals based on a negative binomial null model have also been independently proposed as the sctransform method [60].”
MethodDroplet barcoding for single-cell transcriptomics applied to embryonic stem cells
Klein, A. M., Mazutis, L., Akartuna, I., Tallapragada, N., Veres, A., Li, V., Peshkin, L., Weitz, D. A., & Kirschner, M. W. (2015) · Cell · doi:10.1016/j.cell.2015.04.044 · Cited by 3,356
Finak, G., McDavid, A., Yajima, M., Deng, J., Gersuk, V., Shalek, A. K., Slichter, C. K., Miller, H. W., McElrath, M. J., Prlic, M., et al. (2015) · Genome Biology · doi:10.1186/s13059-015-0844-5 · Cited by 3,159
Deep generative modeling for single-cell transcriptomics
Lopez, R., Regier, J., Cole, M. B., Jordan, M. I., & Yosef, N. (2018) · Nature Methods · doi:10.1038/s41592-018-0229-2 · Cited by 2,530
Smart-seq2 for sensitive full-length transcriptome profiling in single cells
Picelli, S., Björklund, Å. K., Faridani, O. R., Sagasser, S., Winberg, G., & Sandberg, R. (2013) · Nature Methods · doi:10.1038/nmeth.2639 · Cited by 2,358
Isometric Logratio Transformations for Compositional Data Analysis
Egozcue, J. J., Pawlowsky‐Glahn, V., Mateu‐Figueras, G., & Barceló-Vidal, C. (2003) · Mathematical Geology · doi:10.1023/a:1023818214614 · Cited by 1,956
Single-cell mRNA quantification and differential analysis with Census
Qiu, X., Hill, A. J., Packer, J. S., Lin, D., Ma, Y.-A., & Trapnell, C. (2017) · Nature Methods · doi:10.1038/nmeth.4150 · Cited by 1,904
McCarthy, D. J., Campbell, K. R., Lun, A. T. L., & Wills, Q. F. (2017) · Bioinformatics · doi:10.1093/bioinformatics/btw777 · Cited by 1,852
A single-cell survey of the small intestinal epithelium
Haber, A. L., Biton, M., Rogel, N., Herbst, R. H., Shekhar, K., Smillie, C., Burgin, G., Delorey, T. M., Howitt, M. R., Katz, Y., et al. (2017) · Nature · doi:10.1038/nature24489 · Cited by 1,750
A step-by-step workflow for low-level analysis of single-cell RNA-seq data
Lun, A. T. L., McCarthy, D. J., & Marioni, J. C. (2016) · F1000Research · doi:10.12688/f1000research.9501.1 · Cited by 1,551
The Technology and Biology of Single-Cell RNA Sequencing
Kolodziejczyk, A. A., Kim, J. K., Svensson, V., Marioni, J. C., & Teichmann, S. A. (2015) · Molecular Cell · doi:10.1016/j.molcel.2015.04.005 · Cited by 1,487
A Single-Cell Transcriptome Atlas of the Human Pancreas
Muraro, M. J., Dharmadhikari, G., Grün, D., Gröen, N., Dielen, T., Jansen, E., van Gurp, L., Engelse, M. A., Carlotti, F., de Koning, E. J. P., et al. (2016) · Cell Systems · doi:10.1016/j.cels.2016.09.002 · Cited by 1,307
Quantitative single-cell RNA-seq with unique molecular identifiers
Islam, S., Zeisel, A., Joost, S., La Manno, G., Zajac, P., Kasper, M., Lönnerberg, P., & Linnarsson, S. (2014) · Nature Methods · doi:10.1038/nmeth.2772 · Cited by 1,183
Single-cell sequencing-based technologies will revolutionize whole-organism science
Shapiro, E., Biezuner, T., & Linnarsson, S. (2013) · Nature Reviews Genetics · doi:10.1038/nrg3542 · Cited by 1,177
Pooling across cells to normalize single-cell RNA sequencing data with many zero counts
L. Lun, A. T., Bach, K., & Marioni, J. C. (2016) · Genome Biology · doi:10.1186/s13059-016-0947-7 · Cited by 1,108
Exponential scaling of single-cell RNA-seq in the past decade
Svensson, V., Vento-Tormo, R., & Teichmann, S. A. (2018) · Nature Protocols · doi:10.1038/nprot.2017.149 · Cited by 1,034
Accounting for technical noise in single-cell RNA-seq experiments
Brennecke, P., Anders, S., Kim, J. K., Kołodziejczyk, A. A., Zhang, X., Proserpio, V., Baying, B., Benes, V., Teichmann, S. A., Marioni, J. C., et al. (2013) · Nature Methods · doi:10.1038/nmeth.2645 · Cited by 979
Bias, robustness and scalability in single-cell differential expression analysis
Soneson, C., & Robinson, M. D. (2018) · Nature Methods · doi:10.1038/nmeth.4612 · Cited by 770
Algorithm AS136 : A K-means clustering algorithm
Hartigan, J. A. (1979) · Journal of the Royal Statistical Society Series C (Applied Statistics) · Cited by 743
Validation of noise models for single-cell transcriptomics
Grün, D., Kester, L., & van Oudenaarden, A. (2014) · Nature Methods · doi:10.1038/nmeth.2930 · Cited by 721
A general and flexible method for signal extraction from single-cell RNA-seq data
Risso, D., Perraudeau, F., Gribkova, S., Dudoit, S., & Vert, J.-P. (2018) · Nature Communications · doi:10.1038/s41467-017-02554-5 · Cited by 718
SpatialDE: identification of spatially variable genes
Svensson, V., Teichmann, S. A., & Stegle, O. (2018) · Nature Methods · doi:10.1038/nmeth.4636 · Cited by 695
Missing data and technical variability in single-cell RNA-sequencing experiments
Hicks, S. C., Townes, F. W., Teng, M., & Irizarry, R. A. (2018) · Biostatistics · doi:10.1093/biostatistics/kxx053 · Cited by 635
Self-citationMissing Data and Technical Variability in Single-Cell RNA-Sequencing Experiments
Hicks, S. C., Townes, F. W., Teng, M., & Irizarry, R. A. (2015) · doi:10.1101/025528 · Cited by 625 · cited 2×
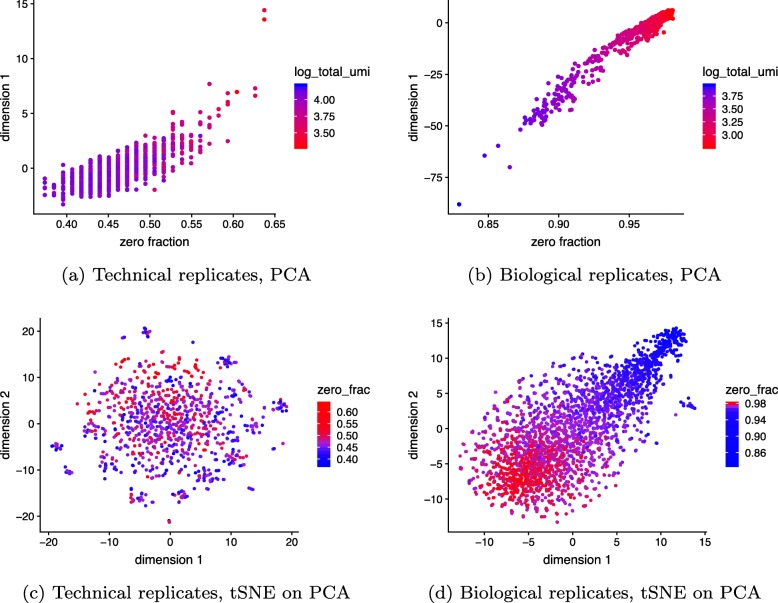
“Focusing on the entire negative control datasets, we applied PCA to the log transformed CPMs and observed a strong correlation between the first principal component (PC) and the fraction of zeros, consistent with [28].”
BackgroundMethodSelf-citationZIFA: Dimensionality reduction for zero-inflated single-cell gene expression analysis
Pierson, E., & Yau, C. (2015) · Genome Biology · doi:10.1186/s13059-015-0805-z · Cited by 577
CIDR: Ultrafast and accurate clustering through imputation for single-cell RNA-seq data
Lin, P., Troup, M., & Ho, J. W. K. (2017) · Genome biology · doi:10.1186/s13059-017-1188-0 · Cited by 541
Foundations of Linear and Generalized Linear Models
Agresti, A. (2015) · CERN Document Server (European Organization for Nuclear Research) · Cited by 473
Normalizing single-cell RNA sequencing data: challenges and opportunities
Vallejos, C. A., Risso, D., Scialdone, A., Dudoit, S., & Marioni, J. C. (2017) · Nature Methods · doi:10.1038/nmeth.4292 · Cited by 436
A Generalization of Principal Components Analysis to the Exponential Family
Collins, M., Dasgupta, S., & Schapire, R. E. (2001) · Cited by 383
Droplet scRNA-seq is not zero-inflated
Svensson, V. (2019) · doi:10.1101/582064 · Cited by 356
Batch effects and the effective design of single-cell gene expression studies
Tung, P.-Y., Blischak, J. D., Hsiao, C. J., Knowles, D. A., Burnett, J. E., Pritchard, J. K., & Gilad, Y. (2017) · Scientific Reports · doi:10.1038/srep39921 · Cited by 356
Kim, J., He, Y., & Park, H. (2014) · Journal of Global Optimization · doi:10.1007/s10898-013-0035-4 · Cited by 352
M3Drop: dropout-based feature selection for scRNASeq
Andrews, T. S., & Hemberg, M. (2018) · Bioinformatics · doi:10.1093/bioinformatics/bty1044 · Cited by 298
A systematic performance evaluation of clustering methods for single-cell RNA-seq data
Duò, A., Robinson, M. D., & Soneson, C. (2018) · F1000Research · doi:10.12688/f1000research.15666.2 · Cited by 296
Identifying cell populations with scRNASeq
Andrews, T. S., & Hemberg, M. (2017) · Molecular Aspects of Medicine · doi:10.1016/j.mam.2017.07.002 · Cited by 247
Hafemeister, C., & Satija, R. (2019) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/576827 · Cited by 238
Single-cell transcriptome sequencing: recent advances and remaining challenges
Liu, S., & Trapnell, C. (2016) · F1000Research · doi:10.12688/f1000research.7223.1 · Cited by 230
Observation weights unlock bulk RNA-seq tools for zero inflation and single-cell applications
Van den Berge, K., Perraudeau, F., Soneson, C., Love, M. I., Risso, D., Vert, J.-P., Robinson, M. D., Dudoit, S., & Clement, L. (2018) · Genome Biology · doi:10.1186/s13059-018-1406-4 · Cited by 228
Classification and clustering of sequencing data using a Poisson model
Witten, D. (2011) · The Annals of Applied Statistics · doi:10.1214/11-aoas493 · Cited by 186
Scalable Recommendation with Poisson FactorizationPreprint
Gopalan, P., Hofman, J. M., & Blei, D. M. (2013) · arXiv (Cornell University) · doi:10.48550/arxiv.1311.1704 · Cited by 176
Models for transcript quantification from RNA-SeqPreprint
Pachter, L. S. (2011) · arXiv (Cornell University) · doi:10.48550/arxiv.1104.3889 · Cited by 175
Naught all zeros in sequence count data are the same
Silverman, J. D., Roche, K. E., Mukherjee, S., & David, L. A. (2020) · Computational and Structural Biotechnology Journal · doi:10.1016/j.csbj.2020.09.014 · Cited by 171
Exponential scaling of single-cell RNA-seq in the last decadePreprint
Svensson, V., Vento‐Tormo, R., & Teichmann, S. A. (2017) · arXiv (Cornell University) · doi:10.48550/arxiv.1704.01379 · Cited by 169
The Multinomial-Poisson Transformation
Baker, S. G. (1994) · Journal of the Royal Statistical Society Series D (The Statistician) · doi:10.2307/2348134 · Cited by 136
Synthetic evolutionary origin of a proofreading reverse transcriptase
Ellefson, J. W., Gollihar, J., Shroff, R., Shivram, H., Iyer, V. R., & Ellington, A. D. (2016) · Science · doi:10.1126/science.aaf5409 · Cited by 135
K-nearest neighbor smoothing for high-throughput single-cell RNA-Seq dataPreprint
Wagner, F., Yan, Y., & Yanai, I. (2017) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/217737 · Cited by 134
How to design a single-cell RNA-sequencing experiment: pitfalls, challenges and perspectives
Molin, A. D., & Camillo, B. D. (2018) · Briefings in Bioinformatics · doi:10.1093/bib/bby007 · Cited by 97
A systematic performance evaluation of clustering methods for single-cell RNA-seq data
Duò, A., Robinson, M. D., & Soneson, C. (2018) · F1000Research · doi:10.12688/f1000research.15666.1 · Cited by 97
Distributed Multinomial Regression
Taddy, M. A. (2016) · Cited by 76
Distributed multinomial regression
Taddy, M. (2015) · The Annals of Applied Statistics · doi:10.1214/15-aoas831 · Cited by 62
Why You Cannot Transform Your Way out of Trouble for Small Counts
Warton, D. I. (2017) · Biometrics · doi:10.1111/biom.12728 · Cited by 57
A brief review of single-cell transcriptomic technologies
Tomer Kalisky, C., Oriel, S., Bar-Lev, T. H., Ben-Haim, N., Trink, A., Wineberg, Y., Kanter, I., Gilad, S., & Pyne, S. (2017) · Briefings in Functional Genomics · doi:10.1093/bfgp/elx019 · Cited by 56
Lun, A. T. L. (2018) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/404962 · Cited by 56
ZINB-WaVE: A general and flexible method for signal extraction from single-cell RNA-seq dataPreprint
Risso, D., Perraudeau, F., Gribkova, S., et al. (2017) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/125112 · Cited by 43
On the poisson approximation to the multinomial distribution
McDonald, D. R. (1980) · Canadian Journal of Statistics · doi:10.2307/3314676 · Cited by 36
Naught all zeros in sequence count data are the samePreprint
Silverman, J. D., Roche, K. E., Mukherjee, S., et al. (2018) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/477794 · Cited by 34
Bayesian Inference for a Generative Model of Transcriptome Profiles from Single-cell RNA SequencingPreprint
Lopez, R., Regier, J., Cole, M., et al. (2018) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/292037 · Cited by 31
A robust nonlinear low-dimensional manifold for single cell RNA-seq data
Verma, A., & Engelhardt, B. E. (2020) · BMC Bioinformatics · doi:10.1186/s12859-020-03625-z · Cited by 29
A general framework for association analysis of heterogeneous data
Li, G., & Gaynanova, I. (2018) · Project Euclid (Cornell University) · doi:10.1214/17-aoas1127 · Cited by 22
Landgraf, A. J. (2015) · OhioLink ETD Center (Ohio Library and Information Network) · Cited by 9
A robust nonlinear low-dimensional manifold for single cell RNA-seq dataPreprint
Verma, A., & Engelhardt, B. E. (2018) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/443044 · Cited by 8
Generalized Principal Component AnalysisPreprint
Townes, F. W. (2019) · arXiv (Cornell University) · doi:10.48550/arxiv.1907.02647 · Cited by 2
Self-citationwilltownes/scrna2019: Genome Biology publication
Townes, W., & Pita-Juarez, Y. (2019) · Zenodo (CERN European Organization for Nuclear Research) · doi:10.5281/zenodo.3475534 · Cited by 1
The latent logarithmPreprint
Biswas, S. (2016) · arXiv (Cornell University) · doi:10.48550/arxiv.1605.06064
Cited by
Sorted by citations
Hafemeister, C., & Satija, R. (2019) · Genome Biology · doi:10.1186/s13059-019-1874-1 · Cited by 4,479
“For example, Townes et al. [29] introduced GLM-PCA, a generalization of PCA for data exhibiting non-normal error distributions such as the negative binomial, that takes count data directly as input instead of relying on intermediate residuals.”
MethodEleven grand challenges in single-cell data science
Lähnemann, D., Köster, J., Szczurek, E., McCarthy, D. J., Hicks, S. C., Robinson, M. D., Vallejos, C. A., Campbell, K. R., Beerenwinkel, N., Mahfouz, A., et al. (2020) · Genome Biology · doi:10.1186/s13059-020-1926-6 · Cited by 1,327
Self-citationRobust decomposition of cell type mixtures in spatial transcriptomics
Cable, D. M., Murray, E., Zou, L. S., Goeva, A., Macosko, E. Z., Chen, F., & Irizarry, R. A. (2022) · Nature Biotechnology · doi:10.1038/s41587-021-00830-w · Cited by 1,487
Self-citationBest practices for single-cell analysis across modalities
Heumos, L., Schaar, A. C., Lance, C., Litinetskaya, A., Drost, F., Zappia, L., Lücken, M. D., Strobl, D. C., Henao, J., Curion, F., et al. (2023) · Nature Reviews Genetics · doi:10.1038/s41576-023-00586-w · Cited by 1,051
“Deviance identifies highly informative genes by fitting a gene-wise model that assumes constant expression across all cells and quantifying which genes violate this assumption 47 .”
BackgroundThe art of using t-SNE for single-cell transcriptomics
Kobak, D., & Berens, P. (2019) · Nature Communications · doi:10.1038/s41467-019-13056-x · Cited by 1,161
Metadata
- Authors
- Townes, F. W., Hicks, S. C., Aryee, M. J., & Irizarry, R. A.
- Affiliations
- Harvard University (US)
- Johns Hopkins University (US)
- Venue
- Genome Biology
- Published
- 2019-12
- Volume
- 20
- Issue
- 1
- Pages
- 295-295
- ISSN
- 1465-6906 (ISSN-L)
- 1474-760X
- PMC
- PMC6927135
- OpenAlex
- W2998065416
- Cited by
- 471
- References
- 68
- Citations by year
- 2026: 49 · 2025: 56 · 2024: 78 · 2023: 76 · 2022: 70 · 2021: 80 · 2020: 48 · 2019: 14
- FWCI
- 17.71
- Type
- journal article
- Language
- en
- Update notices
- correction (2020-07-22)
- Open access
- gold
- doi.org
- Licence
- Locations
- Genome biology (publishedVersion, cc-by)
- bioRxiv (Cold Spring Harbor Laboratory) (acceptedVersion, cc-by-nc-nd)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- DOAJ (DOAJ: Directory of Open Access Journals) (submittedVersion)
- Funders
- National Institutes of Health
- National Human Genome Research Institute (R00HG009007, R01HG005220, P41HG004059)
- National Cancer Institute (T32CA009337)
- National Institute of General Medical Sciences (R01GM083084)
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Single-cell and spatial transcriptomics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Gene expression and cancer classification
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Genomics and Phylogenetic Studies
- Keywords
- single-cell RNA sequencing, feature selection, dimensionality reduction, multinomial model, unique molecular identifiers, highly variable genes, generalized principal component analysis, cell clustering
- MeSH terms
- Models, Statistical, Sequence Analysis, RNA, Single-Cell Analysis
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
