

AlphaFold Protein Structure Database in 2024: providing structure coverage for over 214 million protein sequences
Open accessThe DOI matches its Crossref record; 29 of the 29 references it cites were found in a registry or an index; 18 of the 18 sources its claims cite were checked for support.
Abstract
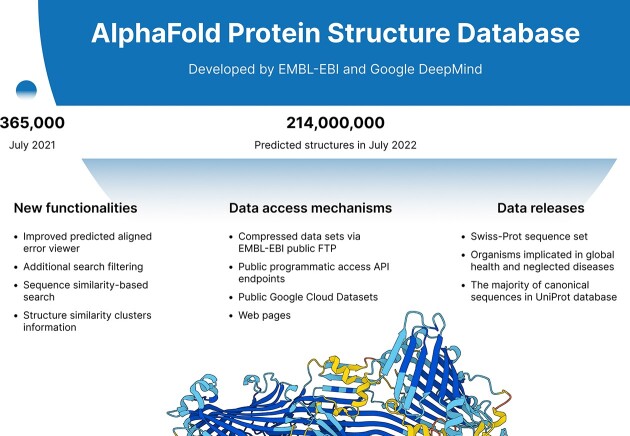
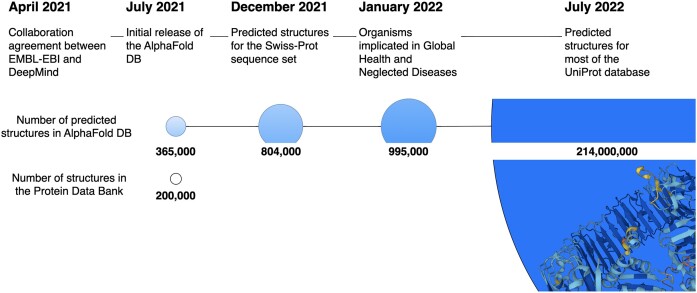
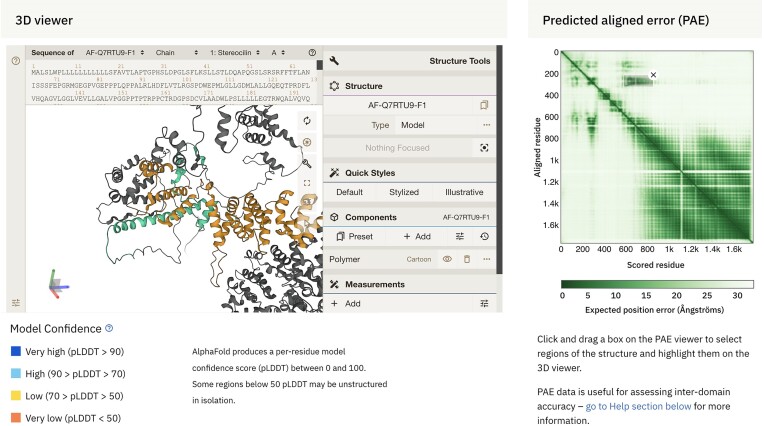
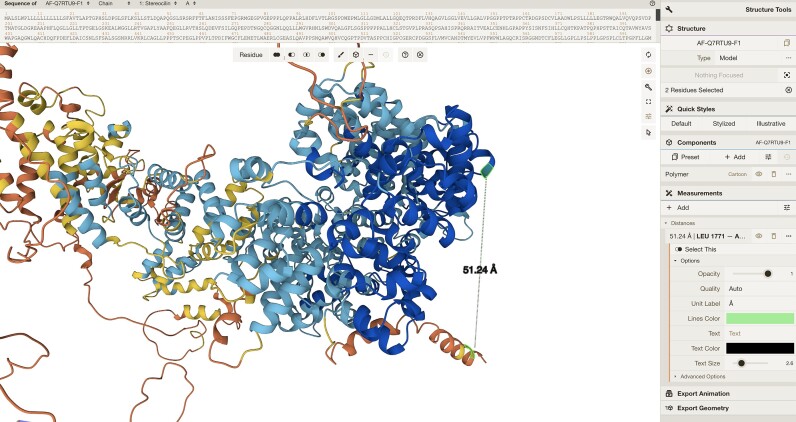
The AlphaFold Database Protein Structure Database (AlphaFold DB, https://alphafold.ebi.ac.uk) has significantly impacted structural biology by amassing over 214 million predicted protein structures, expanding from the initial 300k structures released in 2021. Enabled by the groundbreaking AlphaFold2 artificial intelligence (AI) system, the predictions archived in AlphaFold DB have been integrated into primary data resources such as PDB, UniProt, Ensembl, InterPro and MobiDB. Our manuscript details subsequent enhancements in data archiving, covering successive releases encompassing model organisms, global health proteomes, Swiss-Prot integration, and a host of curated protein datasets. We detail the data access mechanisms of AlphaFold DB, from direct file access via FTP to advanced queries using Google Cloud Public Datasets and the programmatic access endpoints of the database. We also discuss the improvements and services added since its initial release, including enhancements to the Predicted Aligned Error viewer, customisation options for the 3D viewer, and improvements in the search engine of AlphaFold DB.
Overview
CoChat Summary
TL;DR
AlphaFold DB has grown from ~300k to over 214 million predicted protein structures as of September 2023, with expanded access, search, and visualization features. This 2024 update describes data archiving, access mechanisms, integration with core resources, and new clustering/BLAST/PAE viewer capabilities.
Key findings
- AlphaFold DB expanded from approximately 300k structures in 2021 to over 214 million predicted protein structures as of September 2023; the initial July 2021 release housed over 360,000 structures for 20 model organism proteomes.
- Predictions are integrated into resources including PDB, UniProt, Ensembl, InterPro and MobiDB, and are accessible via FTP, Google Cloud Public Datasets (~23 TiB), API, and the website; PAE data is unavailable from EMBL-EBI public FTP.
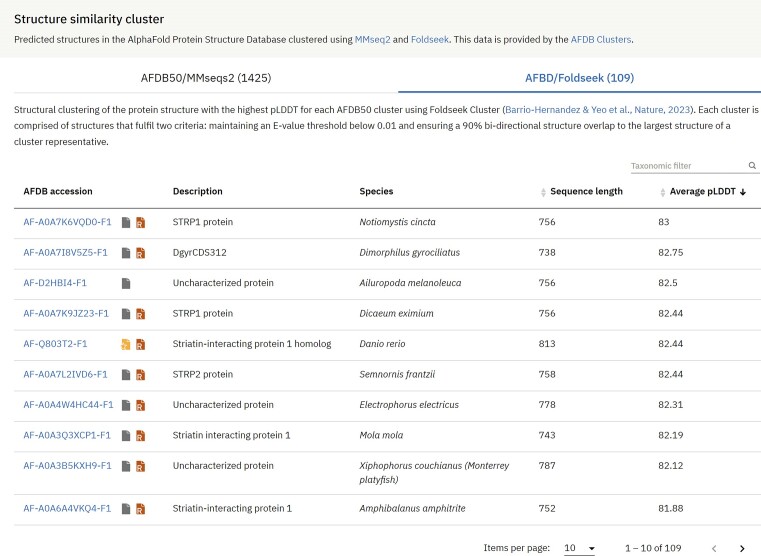
- Structure similarity clustering used MMseqs2 to cluster 214 million UniProtKB sequences into 52 million clusters, then Foldseek secondary clustering yielded 18.8 million clusters, finalizing 2.30 million robust clusters each with at least a pair of structures.
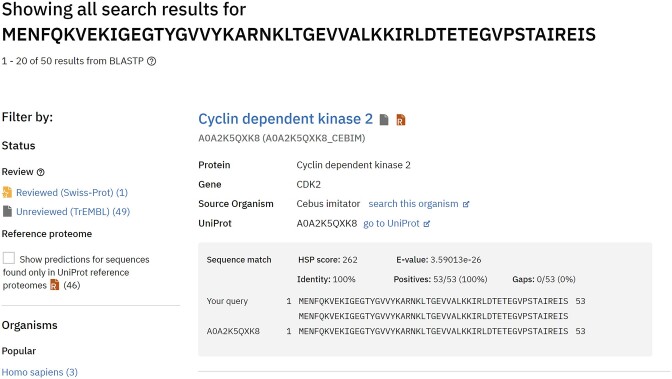
- New features include BLAST sequence similarity search, improved search filters for reviewed/unreviewed and reference proteome status, enhanced PAE viewer–Mol* interactivity, and a November 2022 update fixing a numerical bug that affected ~4% of total structure predictions.
Methods
- Descriptive database update/report with no primary experimental design; data generation is carried out by Google DeepMind, with predictions stored in PDB, mmCIF (modelCIF), and binaryCIF formats plus JSON metadata and confidence metrics pLDDT and PAE.
- Archiving uses successive releases from UniProt reference proteomes (e.g., 2021_02, 2021_04), Swiss-Prot, WHO global health proteomes, and MANE select; as of September 2023, EMBL-EBI FTP hosts TAR files for proteomes of 48 organisms.
- Access and analysis infrastructure includes FTP, Google Cloud Public Datasets, an API keyed on UniProt accessions, Apache-Solr search, BLAST on Google Cloud Platform, and Foldseek/MMseqs2 clustering.
Limitations
- Stated: sequences are not covered if they are less than 16 amino acids; >2700 aa for Swiss-Prot/proteome sequences or >1280 aa for other UniProt sequences; contain non-standard amino acids; are not in the UniProt ‘one sequence per gene’ FASTA file; or are viral proteins; these limitations are under discussion.
- Stated: some sequences may be outdated compared to UniProt due to less frequent AlphaFold DB releases; PAE data is unavailable from EMBL-EBI public FTP; full Google Cloud dataset download requires a Google account and can take roughly 2.5 days at 1 Gbps.
- Evident from the text: all archived structures are predicted models rather than experimental structures, and users must interpret pLDDT/PAE cautiously (e.g., pLDDT <50 likely disordered, 50–70 lower confidence); the database also currently lacks isoform and multimeric predicted structures, which are listed as future work.
Model-written summary. Check it against the paper before relying on it.
Integrity signals
Registry
Cites
Statistics
Figures

Verification
Claims the paper makes
11 Valid · 1 Not valid · 0 Not checked · 28 Own results and interpretations
Every source the paper cites for this claim was checked and supports it.
In the paper
“The new generation of predicted protein structure models has demonstrated remarkable accuracy, which could mitigate the continually widening gap between known protein sequences and experimentally determined protein structures (7).”
§ AlphaFold Protein Structure Database in 2024: providing structure coverage for over 214 million protein sequences > Introduction
What the sources say
- Supportedcites [7] Varadi M., Bordin N., Orengo C., Velankar S. The opportunities and challenges posed by the new generation of deep learning-based protein structure predictors. Curr. Opin. Struct. Biol. 2023; 79: 102543. 36807079 PMC7618866 source
The Introduction says prediction models made a major advance in accuracy and can match or sometimes surpass experimentally determined structures. It also describes a growing gap between known protein sequences and determined structures; the section on new challenges says predicted structures have greatly increased coverage of protein sequence space. These passages support both the accuracy claim and the proposed potential to narrow that gap (page numbers unavailable). (LLM, conf=0.93)
boosted- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“In response to frequent requests from the user community since the AlphaFold DB launch, we have made significant strides to implement a sequence-based similarity search feature. We have incorporated the Basic Local Alignment Search Tool (BLAST) (21) into our Google Cloud Platform infrastructure to achieve this.”
§ AlphaFold Protein Structure Database in 2024: providing structure coverage for over 214 million protein sequences > Implementation > Data access
What the sources say
- Supportedcites [21] Camacho C., Coulouris G., Avagyan V., Ma N., Papadopoulos J., Bealer K., Madden T.L. BLAST+: architecture and applications. BMC Bioinf. 2009; 10: 421. PMC2803857 20003500 source
The Background identifies BLAST as a sequence similarity search program that matches a query sequence against a database. In the citing sentence, the citation identifies the tool incorporated; the AlphaFold DB implementation is the citing authors’ own report, not a claim attributed to this source. (LLM, conf=0.91)
plainstates a cause- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“The impact of the predicted structures is further enhanced by their seamless integration into several core data resources, including PDBe-KB (17), UniProt (6), Ensembl (16), InterPro (15), Genecards (24) and MobiDB (25), among others.”
§ AlphaFold Protein Structure Database in 2024: providing structure coverage for over 214 million protein sequences > Conclusions and outlook
What the sources say
- Supportedcites [17] consortium P.D.B.-K.B. PDBe-KB: collaboratively defining the biological context of structural data. Nucleic Acids Res. 2022; 50: D534– D542. 34755867 PMC8728252 source
Citation (17) is attached specifically to PDBe-KB, rather than to the other resources in the sentence. The Introduction identifies PDBe-KB as a resource that integrates structural and functional annotations, supporting its identification here. (LLM, conf=0.90)
- Supportedcites [6] U.P. Consortium UniProt: the Universal Protein Knowledgebase in 2023. Nucleic Acids Res. 2023; 51: D523– D531. 36408920 PMC9825514 source
The citation is attached to UniProt in a list of resources, and the cited paper is a description of UniProt. Its Data integration and Interactive visualisations in entry pages sections state that UniProt incorporates AlphaFold-predicted structures into protein entries alongside other data; the supplied passages do not give page numbers. (LLM, conf=0.93)
- Supportedcites [16] Cunningham F., Allen J.E., Allen J., Alvarez-Jarreta J., Amode M.R., Armean I.M., Austine-Orimoloye O., Azov A.G., Barnes I., Bennett R. et al. Ensembl 2022. Nucleic Acids Res. 2022; 50: D988– D995. 34791404 PMC8728283 source
The citation identifies Ensembl as one of the resources in the list. In “Developing a new website” (page not provided), the paper says Ensembl’s Entity Viewer provides access to protein information, including 3D structures from PDBe. (LLM, conf=0.86)
plainstates a cause- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“Tackling this challenge, Steinegger et al. recently unveiled the Foldseek Cluster, a state-of-the-art structural-alignment-based clustering algorithm designed for vast datasets (22). They applied this novel method to the AlphaFold DB archive, making the derived structure similarity clusters available to the research community.”
§ AlphaFold Protein Structure Database in 2024: providing structure coverage for over 214 million protein sequences > Implementation > Data access
What the sources say
- Supportedcites [22] Barrio-Hernandez I., Yeo J., Jänes J., Mirdita M., Gilchrist C.L.M., Wein T., Varadi M., Velankar S., Beltrao P., Steinegger M. Clustering-predicted structures at the scale of the known protein universe. Nature. 2023; 622: 637– 645. 37704730 PMC10584675 source
The Abstract says the authors used Foldseek cluster to cluster the AlphaFold database. The Methods section (“Web server”) says they developed a server for users to explore the clusters, their members and related clusters, supporting the claim that the results were made available to researchers. (LLM, conf=0.93)
plainstates a cause- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
- Probably not a substantial claim. 30
- It may state two separate claims as one. 15
- The quote only partly states this claim. 1
Claims other papers make about this one
Each is checked against this paper’s own text: does it say what the citing paper says it does?
The cited source says what the claim attributes to it.
In the citing paper
“houses over 200 million pre-run AF2 predictions”
From The power and pitfalls of AlphaFold2 for structure prediction beyond rigid globular proteins (2024) · cites this paper as [7]
Citations
Cites
Highly accurate protein structure prediction with AlphaFold
Jumper, J., Evans, R., Pritzel, A., Green, T., Figurnov, M., Ronneberger, O., Tunyasuvunakool, K., Bates, R., Žídek, A., Potapenko, A., et al. (2021) · Nature · doi:10.1038/s41586-021-03819-2 · Cited by 43,764
Self-citationBLAST+: architecture and applications
Camacho, C., Coulouris, G., Avagyan, V., Ma, N., Papadopoulos, J., Bealer, K., & Madden, T. L. (2009) · BMC Bioinformatics · doi:10.1186/1471-2105-10-421 · Cited by 22,074
UniProt: the Universal Protein Knowledgebase in 2023
The UniProt Consortium, Bateman, A., Martin, M.-J., Orchard, S., Magrane, M., Ahmad, S., Alpi, E., Bowler-Barnett, E. H., Britto, R., Bye-A-Jee, H., et al. (2023) · Nucleic Acids Research · doi:10.1093/nar/gkac1052 · Cited by 7,210
Evolutionary-scale prediction of atomic level protein structure with a language model
Lin, Z., Akin, H., Rao, R., Hie, B., Zhu, Z., Lu, W., Smetanin, N., Verkuil, R., Kabeli, O., Shmueli, Y., et al. (2022) · doi:10.1101/2022.07.20.500902 · Cited by 5,990 · cited 1×
“In the wake of the latest breakthroughs in prediction software, new databases such as the AlphaFold Protein Structure Database and the ESM Atlas ( 14 ) have emerged.”
MethodThe GeneCards suite: from gene data mining to disease genome sequence analyses
Stelzer, G., Rosen, N., Plaschkes, I., Zimmerman, S., Twik, M., Fishilevich, S., Stein, T. I., Nudel, R., Lieder, I., Mazor, Y., et al. (2016) · Current Protocols in Bioinformatics · doi:10.1002/cpbi.5 · Cited by 5,550 · cited 1×
“The impact of the predicted structures is further enhanced by their seamless integration into several core data resources, including PDBe-KB ( 17 ), UniProt ( 6 ), Ensembl ( 16 ), InterPro ( 15 ), Genecards ( 24 ) and MobiDB ( 25 ), among others.”
BackgroundAccurate prediction of protein structures and interactions using a 3-track network
Baek, M., DiMaio, F., Anishchenko, I., Dauparas, J., Ovchinnikov, S., Lee, G. R., Wang, J., Cong, Q., Kinch, L. N., Schaeffer, R. D., et al. (2021) · doi:10.1101/2021.06.14.448402 · Cited by 5,549
Paysan-Lafosse, T., Blum, M., Chuguransky, S., Grego, T., Pinto, B. L., Salazar, G. A., Bileschi, M. L., Bork, P., Bridge, A., Colwell, L., et al. (2023) · Nucleic Acids Research · doi:10.1093/nar/gkac993 · Cited by 2,399
Cunningham, F., Allen, J. E., Allen, J., Alvarez-Jarreta, J., Amode, M. R., Armean, I. M., Austine-Orimoloye, O., Azov, A. G., Barnes, I., Bennett, R., et al. (2022) · Nucleic Acids Research · doi:10.1093/nar/gkab1049 · Cited by 1,923
Mol* Viewer: modern web app for 3D visualization and analysis of large biomolecular structures
Sehnal, D., Bittrich, S., Deshpande, M., Svobodová, R., Berka, K., Bazgier, V., Velankar, S., Burley, S. K., Koča, J., & Rose, A. S. (2021) · Nucleic Acids Research · doi:10.1093/nar/gkab314 · Cited by 1,154 · cited 1×
“We improved the visualisation of non-consecutive regions and the interactivity between the PAE viewer and the 3D molecular graphics viewer, Mol* ( 23 ) (Figure 5 ).”
MethodSelf-citationMariani, V., Biasini, M., Barbato, A., & Schwede, T. (2013) · Bioinformatics · doi:10.1093/bioinformatics/btt473 · Cited by 1,138 · cited 1×
“AlphaFold yields a per-residue estimate of its confidence, known as pLDDT, ranging from 0 to 100, indicating the tool’s predicted score on the lDDT-C α metric ( 19 ).”
Method
Cited by
Items 1,101–1,200 of 2,174
Cryo-EM structure of the human COP1-DET1 ubiquitin ligase complex
Wang, S., Teng, F., Stjepanović, G., Rao, F., & Su, M.-Y. (2026) · Nature Communications · doi:10.1038/s41467-026-68375-7 · Cited by 3
“…further performed particle subtraction on this domain to improve the resolution to 2.65Å, and the resulted reconstruction enabled the fi tting of the individual atomic models for DDB1, DDA1 (PDB: 6DSZ) and DET1 (AlphaFold model: AF-Q7L5Y6-F1), allowing for the modelling of most of the complex 34 .”
Liang, J., Tang, X., Li, H., Fang, Y., & Jiang, F. (2026) · Journal of Peptide Science · doi:10.1002/psc.70119 · Cited by 2
“…structure prediction models represented by AlphaFold3 can now predict the three-dimensional structures of peptides with atomic-level precision [152–154], whereas large language model-based tools such as Evolutionary Scale Modeling Fold (ESMFold) have increased prediction speeds by orders of…”
Rakshit, R., Bahl, A., Gautam, G., et al. (2025) · Journal of Molecular Graphics and Modelling · doi:10.1016/j.jmgm.2025.109185 · Cited by 1
ASMC: investigating the amino acid diversity of enzyme active sites
Bailly, T., Elisée, E., & Vallenet, D. (2025) · Bioinformatics · doi:10.1093/bioinformatics/btaf307 · Cited by 1
“This step can be skipped if the user already has a set of 3D structures, e.g. taken from AI-based databases (Lin et al. 2023, Varadi et al. 2024), though care must be taken to ensure pocket accuracy for reliable active site identification and comparison.”
Foudah, A. I., Alqarni, M. H., Aljarba, T. M., Jawaid, T., Alkhamees, O. A., Alsanad, S. M., AlHussaini, K. I., & Alam, A. (2025) · Molecular Diversity · doi:10.1007/s11030-025-11339-8 · Cited by 1
“The recent advent of AlphaFold, a deep learning-based protein structure prediction tool, now enables high-confidence modeling of previously unresolved targets, thereby expanding the horizon for rational drug discovery [31–33].”
Samak, N. A., Adjir, K., Götz, F., et al. (2026) · Applied and Environmental Microbiology · doi:10.1128/aem.02248-25 · Cited by 1
Large Numbers of New Human Paralogs DiscoveredPreprint
Pradeep, B. K., Deng, W., Hemmat, M. A., Daniels, I., & Jernigan, R. L. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.10.15.680306 · Cited by 1
“Of these, 20,208 proteins have predicted structures available in the AlphaFold Database, 4,25 enabling structural comparisons for almost all pairs, which we use for validating the new paralogs.”
Bouras, G. S., Lim, S. W., Durr, L., Vreugde, S., Goesmann, A., Edwards, R. A., & Schwengers, O. (2026) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.64898/2026.03.31.715528 · Cited by 5
Selçuk, B., Schulte, G., Zhulin, I. B., & Adebalı, O. (2026) · British Journal of Pharmacology · doi:10.1111/bph.70538 · Cited by 1
Wang, H., Guo, Y., Ji, Y., Liu, Q., Chen, Q., Zhang, Q., Yang, M., Valverde, B. E., & Chen, S. (2025) · Pest Management Science · doi:10.1002/ps.70469 · Cited by 2
“42,43 AlphaFold 3 was used to model the position of IVA within the Q B of the A. adenophora D1 protein, revealing that IVA forms a key hydrogen bond (2.21 Å) between the hydroxyl oxygen atom at its 3-position and the residue D1-His215 (Fig.”
InfluentialAltered NaV1.9 channel activity in two Tyr66Ser variant carriers with small fiber dysfunction
van den Braak, N. W. M., Kuehs, S., Peschke, G. Z., et al. (2025) · The Journal of General Physiology · doi:10.1085/jgp.202413691 · Cited by 1
Daniels, P. W., Kelly, S., Tebbs, I. J., & Mitchell, P. (2025) · PLOS One · doi:10.1371/journal.pone.0321120 · Cited by 1
A High-Performance Scientific Database Supporting In-situ Data Query and Accessing
Peng, L., & Chen, Z. (2025) · Asia Conference on Algorithms, Computing and Machine Learning (CACML) · doi:10.1109/cacml64929.2025.11010937 · Cited by 1
“The well-known AlphaFold is trained from a large proteome database[1].”
Kishida, K., Kudo, K., Kumagai, R., Ijima, S., Deng, W., Stari, L., Ogawa-Kishida, N., Ohtsubo, Y., Nagata, Y., & Tsuda, M. (2025) · mBio · doi:10.1128/mbio.01600-25 · Cited by 1
“To obtain predicted structures of these Par proteins, we utilized the AlphaFold Database (44) and AlphaFold Server (45) for structure prediction.”
MPBuild: An Automated Pipeline for High-Fidelity Membrane Protein Simulation System Construction
Liu, Q., Ding, W., Yao, X., Ling, S., & Tian, C. (2025) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.5c01078 · Cited by 1
Maxeiner, S., Walter, L., Zeitz, S. L., & Krasteva‐Christ, G. (2025) · Biology · doi:10.3390/biology14030239 · Cited by 2
Bai, S., Hou, X., Bai, B., et al. (2025) · International Journal of Biological Macromolecules · doi:10.1016/j.ijbiomac.2025.141707 · Cited by 1
Zhang, Y., Li, K., Chen, Y., Zhu, W., Zhang, J., Luo, P., & Lin, A. (2025) · Current Proteomics · doi:10.1016/j.curpro.2025.100053 · Cited by 2
Structural basis of an EMC:Spf1 insertase-dislocase complex in the eukaryotic endoplasmic reticulumPreprint
Klose, C. J., Prabu, J. R., Fenech, E. J., Bräuning, B., Baydar, I., Steigenberger, B., von Gronau, S., Arad, S., Langlois, C. R., Schuldiner, M., et al. (2025) · bioRxiv · doi:10.1101/2025.08.07.669073
“The 3.8 Å reconstruction map of the ATP γ S-bound complex revealed well-defined main-chain and side-chain densities, enabling the fitting of a previously determined structure for EMC (PDB: 7KRA) and a model for Spf1 predicted by AlphaFold 72,73 (AF-P39986-F1).”
InfluentialWang, Z., & Nie, T. (2025) · iScience · doi:10.1016/j.isci.2025.112305 · Cited by 2
“14 While sequence-based methods provide computational efficiency, structural data improves precision, offering deeper insights into protein dynamics and complex binding interactions.”
RBM45 Preferential Binding to m6A: Simulations Suggest Synergy of RRM3 and Other Domains
Park, R., Contreras, L. M., & Tamamis, P. (2026) · The Journal of Physical Chemistry B · doi:10.1021/acs.jpcb.6c01856 · Cited by 1
Clustering intramolecular interactions of proteins significantly enhances cellular thermal stability
Li, M., Chen, X., Qi, H., Tang, Q., Ye, Y., Xiao, X., Zhao, W., Hong, L., & Liu, Z. (2025) · International Journal of Biological Macromolecules · doi:10.1016/j.ijbiomac.2025.148690 · Cited by 1
Banerjee, S., Banerjee, A., Ray, S., Ray, A., Paul, D., Dastidar, S. G., Willard, B. B., & Biswas, K. (2026) · The FASEB Journal · doi:10.1096/fj.202502746r · Cited by 1
“The three-dimensional structure of SP1 was obtained from the AlphaFold Protein Structure Database [32, 33], which web-resource is known to reliably predict models of proteins in an all-atom description.”
Targeting microbial β-CAs: bridging in silico screening with in vitro validation
Tondo, A. R., Fantacuzzi, M., Carradori, S., D’Agostino, I., Angeli, A., Supuran, C. T., Capasso, C., Gambacorta, N., Piemontese, L., Laghezza, A., et al. (2025) · Bioorganic & Medicinal Chemistry · doi:10.1016/j.bmc.2025.118542 · Cited by 1
Öster, C., Chevelkov, V., & Lange, A. (2024) · doi:10.26434/chemrxiv-2024-f6qmr · Cited by 5
Genome mining with hypothetical proteins
Kenney, G. E. (2025) · Methods in enzymology on CD-ROM/Methods in enzymology · doi:10.1016/bs.mie.2025.04.004 · Cited by 1
Catoiu, E. A., Kambo, D., Rodriguez, B., Vear, K., Wang, Z., Akpata, J., Shah, J., Ayoub, R., Hofmann, M., Ananda, K., et al. (2025) · doi:10.1101/2025.09.10.675416 · Cited by 4
Talotta, R., Cammaroto, G., Ruggeri, R. M., Postorino, E. I., Cannavò, S., & Aragona, P. (2025) · International Journal of Molecular Sciences · doi:10.3390/ijms27010200 · Cited by 1
“High scores ( ≥ 90) indicate high confidence of the predicted structure [125].”
He, B., Ali, I., Dorner, L. P., Chowdhury, M. R., Sokolowski, Y., Abadzhieva, E., Reich, A., Wittmaack, M., Sattler, J. M., Frischknecht, F., et al. (2026) · The EMBO Journal · doi:10.1038/s44318-026-00836-7 · Cited by 2
“…SFA2 is similar in size to SFA proteins in other species, consisting of a short N-terminal head domain and a rod domain that has the characteristic repeat regions of the SFA family predicted to form an α -helical structure by AlphaFold (Jumper et al, 2021; Lechtreck, 2003; Varadi et al, 2023).”
Stouthamer, J., Pereira, D., Martin-Ramirez, S., Mutte, S., Mott, G. A., & Smakowska-Luzan, E. (2025) · doi:10.1101/2025.11.26.690834 · Cited by 2
Tayac, C., Torres, J., & Rodas-Rodríguez, J. M. (2025) · SSRN Electronic Journal · doi:10.2139/ssrn.5895214 · Cited by 1
Heckmann, S., Feng, C., Lorenz, J., Dreissig, S., Schubert, V., Wang, B., Hartmann, F., Cuacos, M., Fernández-Jiménez, N., Câmara, A. S., et al. (2024) · doi:10.21203/rs.3.rs-5394998/v1 · Cited by 12
Peptides Unplugged: Small Chains Tackling Big Challenges—One Life-Saving Sequence at a Time!
Bagwe, P. V., Sawant, S. S., & Joshi, S. V. (2025) · International Journal of Peptide Research and Therapeutics · doi:10.1007/s10989-025-10729-8 · Cited by 2
PLM-eXplain: divide and conquer the protein embedding space
van Eck, J., Gogishvili, D., Silva, W., & Abeln, S. (2025) · Bioinformatics · doi:10.1093/bioinformatics/btaf631 · Cited by 7
“We adapted our model using 542 378 Swiss-Prot protein structures from AlphaFoldDB (accessed 18 March 2025), filtered to 25% sequence similarity (Jumper et al. 2021, Varadi et al. 2024).”
Chapman, M. J., Huskey, J. B., Armijo, K. S., Hahn, S., Spellman-Reliford, J., Coleman, A. K., Mabry, C. J., Carpenter, S., Watson, R. O., & Patrick, K. L. (2026) · Infection and Immunity · doi:10.1128/iai.00732-25 · Cited by 1
“Predicted structures for isoforms were obtained from the AlphaFold Protein Structure Database (115, 116), and .pdb files were visualized for annotation using the Pymol software (v2.”
Predicting the Structure of Hydrogenase in Microalgae: The Case of Nannochloropsis salina
Botticelli, S., Faraloni, C., & Penna, G. L. (2025) · Hydrogen · doi:10.3390/hydrogen6040077 · Cited by 1
“For instance AlphaFold Data-base Foldseek [28] is a method ranking structures not only by confidence, but also by structural similarity to all available entries in the AlphaFold data-base.”
Unusual inheritance of a functional cki homolog in the human pathogen Schistosoma mansoni
Wendt, G. R., & Collins, J. J. (2025) · Science Advances · doi:10.1126/sciadv.aea4905 · Cited by 2
Akan, G., Balcı, M. C., Tuncel, G., Karaca, M., Kazan, H. H., Ozketen, A. C., Özgen, Ö., Gökçay, G., & Atalar, F. (2025) · Orphanet Journal of Rare Diseases · doi:10.1186/s13023-025-03775-4 · Cited by 1
“Missense3D utilized AlphaFold models [38] of the proteins obtained from UniProt database.”
Medvedev, K. E., Schaeffer, R. D., & Grishin, N. V. (2025) · Computational and Structural Biotechnology Journal · doi:10.1016/j.csbj.2025.09.018 · Cited by 1
“Recently, The Encyclopedia of Domains (TED) [9] was released - a comprehensive resource for the identification and classification of protein domains within the AlphaFold Database [10].”
Stanley, H. M., & Trent, M. S. (2026) · mBio · doi:10.1128/mbio.03681-25 · Cited by 1
Su, W., Gu, H., Zheng, J., et al. (2025) · Frontiers in Cellular and Infection Microbiology · doi:10.3389/fcimb.2025.1642716 · Cited by 1
Tenger‐Trolander, A., Amiri, E., Gantz, V. M., Kwan, C. W., Yadav, H., Sanders, S. A., & Schmidt‐Ott, U. (2025) · Development · doi:10.1242/dev.204732 · Cited by 2
“…a predicted template modeling (pTM) score>0.5 we then uploaded the structure to FoldSeek ’ s website and searched the available databases (AlphaFold/Proteome, AlphaFold/Swiss-Prot, AlphaFold/ UniProt50, BFMD) for proteins with similar structure (van Kempen et al., 2024; Varadi et al., 2022, 2024).”
InfluentialPrajapati, M., Phogat, S., Jaiwal, R., Jaiwal, P. K., Kumar, L., Ahlawat, Y. K., & Chaudhary, D. (2025) · Molecular Biotechnology · doi:10.1007/s12033-025-01537-y · Cited by 1
“The PDB files of these TLRs were obtained from the AlphaFold Database (TLR3-Q5TJ59, TLR7-A1XC24, TLR4-Q9GL65) ( https:// alpha fold. ebi. ac. uk/) [56] as the target proteins were used for checking the binding affinity with the selected 3D protein construct.”
A pbpB1 mutation causing β-lactam resistance in clinical Listeria monocytogenes isolatesPreprint
Wamp, S., Wagner, R., Schuler, F., et al. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.04.22.649998 · Cited by 1
Absolute hand determination of glycofibrils from natural sources in cryo-EM
Zhang, Q., Qin, L., Wang, T., Li, Z., Zhang, Y., Chen, S., Ge, Q., Wen, J., Yan, N., Wang, J., et al. (2026) · Proceedings of the National Academy of Sciences · doi:10.1073/pnas.2531477123 · Cited by 3
Structural insights into mechanisms of zinc scavenging by the Candida albicans zincophore Pra1Preprint
Syrjänen, J. L., Nore, A., Roselletti, E., Chakraborty, T., Perera, R. L., & Wilson, D. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.01.09.632233 · Cited by 1
Albillos‐Arenal, S., Alonso‐del‐Real, J., Lairón‐Peris, M., et al. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.09.23.614527 · Cited by 1
Bhatnagar, K., Shandilay, S., Malaisamy, A., Bhavesh, N. S., & Chaudhary, S. (2026) · Biochemical and Biophysical Research Communications · doi:10.1016/j.bbrc.2026.153660 · Cited by 1
Computational Tools for Structural Analysis of Proteins
Brezovský, J., Sethi, A., & Surpeta, B. (2024) · Elsevier eBooks · doi:10.1016/b978-0-323-95502-7.00089-0 · Cited by 1
Surbhi, Shau, A. K., Chacko, F. M., & Radhakrishnan, S. K. (2025) · Current Biology · doi:10.1016/j.cub.2025.09.042 · Cited by 1
Sun, G., Fan, Y., Zhu, Y., & Zheng, M. (2025) · BMC Bioinformatics · doi:10.1186/s12859-025-06328-5 · Cited by 1
“It is worth noting that the advent of AlphaFold has provided credible solutions for protein structure prediction [20, 21].”
Auto-crosslinking sporesilk fibers promote endospore and Cry toxin clustering
Remaut, H., Sleutel, M., & Sogues, A. (2025) · doi:10.21203/rs.3.rs-6239967/v1 · Cited by 3
Absolute hand determination of glycofibrils from natural sources in cryo-EM
Zhang, Q., Qin, L., Wang, T., Li, Z., Zhang, Y., Chen, S., Yan, N., Wang, J., & Hu, M. (2025) · doi:10.1101/2025.09.30.679555 · Cited by 3
Much ado about nothing: modeling amino acid replacement with predicted protein structures
(2026) · doi:10.1093/bioinformatics/btag188 · Cited by 1
“All of these matrices predate the arrival of AlphaFold (Jumper et al. 2021; Abramson et al. 2024) and the availability of hundreds of millions of precomputed predicted protein structures (Varadi et al. 2024).”
Crystal structure of exfoliative toxin D from Staphylococcus aureus
Sato, S., Ragab, R. F., Guo, X., et al. (2024) · Biochemical and Biophysical Research Communications · doi:10.1016/j.bbrc.2024.150689 · Cited by 1
Puckelwaldt, O., Gramberg, S., Ajmera, S., Koepke, J., Shamsara, J., Samakovlis, C., Kolb, P., & Haeberlein, S. (2026) · iScience · doi:10.1016/j.isci.2026.116778 · Cited by 4
Xie, X., Shin, Y. H., Kim, T.-S., Seong, Y., Yi, Y. W., & Kim, D. J. (2025) · American Journal of Cancer Research · doi:10.62347/rgzg2120 · Cited by 3
Testing the reliability of AI-generated protein structuresPreprint
Xu, A., & Salzberg, S. L. (2026) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.64898/2026.06.11.731682 · Cited by 1
Shabo, I., Nordling, E., & Abraham‐Nordling, M. (2025) · Biochemistry and Biophysics Reports · doi:10.1016/j.bbrep.2025.102024 · Cited by 1
A diverse interactome of the molecular chaperone CCT/TRiC monomers
Córdoba-Beldad, C. M., & Grantham, J. (2025) · European Journal of Cell Biology · doi:10.1016/j.ejcb.2025.151506 · Cited by 1
BC-Design: A Biochemistry-Aware Framework for Inverse Protein DesignPreprint
Tang, X., Ye, X., Wu, F., Liu, Y., Su, A., Panescu, A., Li, G., Shao, D., Xu, D., & Gerstein, M. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.10.28.620755 · Cited by 2
Saadat, A., & Fellay, J. (2025) · iScience · doi:10.1016/j.isci.2025.112812 · Cited by 4
A global survey of intramolecular isopeptide bonds
Costa, F., Riziotis, I. G., Andreeva, A. E., et al. (2025) · Protein Science · doi:10.1002/pro.70342 · Cited by 1
Decision Tree for Protein Biomarker Selection for Clinical Applications
Waury, K. (2024) · Methods in molecular biology · doi:10.1007/978-1-0716-4298-6_21 · Cited by 1
AlphaFold-Based Protein-Protein Interaction Prediction Methods Classification
Faisal, T., Alofairi, A. A., & Mohsen, . A. (2025) · International Conference on Emerging Smart Technologies and Applications (eSmarTA) · doi:10.1109/esmarta66764.2025.11132127 · Cited by 2
“Structural databases, such as the Protein Data Bank (PDB) and the AlphaFold Database (AFDB), offer detailed insights into the molecular mechanisms underlying protein interactions by providing high-resolution structural data [32, 35].”
Son, H., Lee, S., Kim, J., Park, H., Hwang, M.-H., & Yi, G. (2024) · BMC Bioinformatics · doi:10.1186/s12859-024-05968-3 · Cited by 3
“Protein 3D structural data were obtained in PDB format from the AlphaFold2 database [30], and compound structures were converted from SMILES to SDF format via RDKit.”
MethodBacterial immune systems as causes and consequences of microbiome structure
Custódio, R., Alseth, E., Brockhurst, M. A., Brown, S. P., & Westra, E. R. (2025) · PLoS Biology · doi:10.1371/journal.pbio.3003489 · Cited by 2
Zhang, L., Ma, Y., Hu, J., Liu, F., Jia, Y., Chen, P., Oo, A., Dong, Z., & Pan, M. (2026) · Pesticide Biochemistry and Physiology · doi:10.1016/j.pestbp.2026.107031 · Cited by 1
Guha, S., Bledsoe, R. B., Sutherland, J., et al. (2026) · The ISME Journal · doi:10.1093/ismejo/wrag005 · Cited by 1
“…AlphaFold-Multimer and Foldseek To obtain additional information about the potential for the protein productsofthecandidaterhizobialgenestointeractwithhostproteins such as the NCR peptides directly, we performed in silico protein– protein interaction predictions using AlphaFold-Multimer [54, 55].”
Ligand-Conditioned Side Chain Packing for Flexible Molecular Docking
Luo, D., Qu, X., Lu, D., Wang, Y., Dong, L., & Wang, B. (2025) · Journal of Chemical Theory and Computation · doi:10.1021/acs.jctc.4c01636 · Cited by 3
Fianco, G., Taddei, I., Oropallo, V., Contadini, C., Ferri, A., Coculo, L., Serapian, S. A., Colombo, G., Stagni, V., & Barilà, D. (2026) · Biochimica et Biophysica Acta (BBA) - Molecular Cell Research · doi:10.1016/j.bbamcr.2026.120110 · Cited by 1
1,2,4-Triazole-Based First-in-Class Non-Nucleoside Inhibitors of the Bacterial Enzyme MraY
Berida, T., Huang, T.-Y., Weck, S. C., Lutz, M., McKee, S. R., Kagerah, N., Manning, D. L., Jahan, M. E., Mishra, S., Herrmann, J., et al. (2025) · ACS Bio & Med Chem Au · doi:10.1021/acsbiomedchemau.5c00158 · Cited by 2
Gohain, B. B., Rajkhowa, S., Mazumder, B., et al. (2026) · Molecular Diversity · doi:10.1007/s11030-026-11639-7 · Cited by 1
Song, Y., Hokken, M., Zoll, J., Venselaar, H., Verweij, P. E., Melchers, W. J. G., & Rhodes, J. L. (2026) · Emerging Microbes & Infections · doi:10.1080/22221751.2026.2627078 · Cited by 1
“In addition, Mre11 structures were constructed using AlphaFold for A. fumigatus Mre11 wild-type and Mre11 F332L [19,20].”
Bocquin, L., Reese, F., Schnutenhaus, M., & Hammer, S. C. (2025) · Methods in enzymology on CD-ROM/Methods in enzymology · doi:10.1016/bs.mie.2025.08.020 · Cited by 2
NAC promotes co-translational protein folding at the ribosomal tunnel exitPreprint
Santos, J., Günnigmann, M., Gora, R. J., et al. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.08.02.668148 · Cited by 1
Nasibov, T. F., Gorokhova, A. V., Бразовский, К. С., Ryzhkova, A., Porokhova, E. D., Avdeeva, E. Y., Белоусов, М. В., Kokorev, O. V., & Хлусов, И. А. (2025) · Pharmaceutical Research · doi:10.1007/s11095-025-03934-z · Cited by 1
“For each predicted amino acid residue, a pLDDT (predicted Local Distance Difference Test) confidence score [50, 51] is provided, ranging from 0 to 100.”
Borrajo, M., Greguske, E. A., Maroto, A. F., Palou, A., Renner, A., Giménez-Esbrí, V., Sedano, D., Gut, M. O., Esteve‐Codina, A., Martín‐Mur, B., et al. (2025) · Journal of Biomedical Science · doi:10.1186/s12929-025-01180-4 · Cited by 2
“Automatic annotation in UniProt predicts a signal peptide and Alpha-fold [19, 61] predicts a cell-adhesion protein with five immunoglobulin domains, one fibronectin-type III domain, and one transmembrane domain.”
In vitro reconstitution of minimal human centrosomesPreprint
Rios, M. U., Stachera, W. E., Familiari, N. E., Brito, C., Surrey, T., & Woodruff, J. B. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.02.20.639226 · Cited by 1
DRMAAtic: dramatically improve your cluster potential
Conte, A. D., Ghafouri, H., Clementel, D., et al. (2024) · Bioinformatics Advances · doi:10.1093/bioadv/vbaf112 · Cited by 1
“The UI allows users to upload a protein structure or structural ensemble for analysis or retrieve it from Protein Data Bank (Berman et al. 2003) and AlphaFoldDB (Varadi et al. 2024).”
Xiao, T., Wang, C., Zhu, X., Wang, X., Fan, Y., Jia, X., Lv, J., Chen, T., & Hao, Z. N. (2026) · Veterinary Research · doi:10.1186/s13567-025-01687-6 · Cited by 2
“Homologous structural information for the long tail fiber adhesin and the OmpC were retrieved from SWISS-MODEL [21] and the AlphaFold Protein Structure Database [22] (Additional file 3).”
GENÇ, T. T., Kaya, S., & Günay, M. (2026) · Bulletin of Entomological Research · doi:10.1017/s000748532510076x · Cited by 2
Guntur, G., Gleeson, D., Anderson, M., et al. (2025) · RSC Medicinal Chemistry · doi:10.1039/d5md00353a · Cited by 1
Fang, S., Xu, C., Zhang, J., et al. (2026) · Veterinary Microbiology · doi:10.1016/j.vetmic.2026.110929 · Cited by 1
CK2 directly controls CARD9 protein homeostasisPreprint
Kelly, A. M., Merselis, L. C., Causton, B., Calder, O. K., Chao, H., Prack, M. M., Stine, L. B., Amako, Y., Wang, T., Yesilkanal, A. E., et al. (2025) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2025.02.03.636288 · Cited by 1
Proteome structuring of crown-of-thorns starfish
Zhu, Y., & Lu, Z. (2024) · Frontiers in Marine Science · doi:10.3389/fmars.2024.1487904 · Cited by 1
Deep learning-assisted tools to understand the structural biology of the synapse
Gáspári, Z., Kálmán, Z. E., & Sánta, A. (2025) · Biomedical Engineering Letters · doi:10.1007/s13534-025-00512-5 · Cited by 1
“Now, AlphaFold models are available in the PDB [27], in UniProt [28] and in the huge AlphaFold Protein Structure Database (AlphaFoldDB) with over 200 million entries [29].”
Niroula, D., Charkowski, A. O., Paudel, D., & Joshi, J. R. (2025) · Phytopathology · doi:10.1094/phyto-04-25-0147-r · Cited by 2
Hu, H., Maeshima, H., Tsukahara, R., et al. (2026) · Applied and Environmental Microbiology · doi:10.1128/aem.00727-26 · Cited by 1
ApoDock: Ligand-Conditioned Sidechain Packing for Flexible Molecular DockingPreprint
Luo, D., Qu, X., Lu, D., Wang, Y., Dong, L., & Wang, B. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.11.22.624942 · Cited by 1
Thrift, W. J., Broadwell, Q., Perera, J. F., Lounsbury, N., Chen, J., & Jhunjhunwala, S. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.10.14.618255 · Cited by 3
Got milk? Enzyme-linked immunosorbent assay analysis of casein proteins in methylprednisolone
Hatcher, V. R., Caballero, M. Y., Schuldt, M. M., et al. (2025) · Allergy and Asthma Proceedings · doi:10.2500/aap.2025.46.250047 · Cited by 1
Ninein isoform contributions to intracellular processes and macrophage immune function
Omer, S., Persaud, E., Mohammad, S., Ayo-Farinloye, B., Heineman, R. E.-S., Wellwood, E., Mott, G. A., & Harrison, R. E. (2025) · Journal of Biological Chemistry · doi:10.1016/j.jbc.2025.108419 · Cited by 1
“3.2 (75, 76) via the COSMIC2 platform (77) was used to predict the structures of ninein CAN and ninein ISO2 isoforms.”
InfluentialPPI-ID: Streamlining protein-protein interaction prediction through domain and SLiM mapping
Goodwin, H. V., & Atkinson, N. S. (2025) · PLoS Computational Biology · doi:10.1371/journal.pcbi.1013062 · Cited by 4
“A low error within interdomain quadrants of PAE plots, which are quadrants 1 and 3 (top right and bottom left quadrants; Fig 4), suggests a physical interaction between the proteins [20].”
Karagöl, T., Karagöl, A., & Zhang, S. (2026) · Journal of Molecular Evolution · doi:10.1007/s00239-026-10315-6 · Cited by 4
“Per-residue secondary structure assignments for the studied proteins were obtained from AlphaFold Database (Varadi et al. 2024) models, with the algorithm for D efining the S econdary S tructure of P ro-teins (DSSP) run through the xssp website ( h t t p s : / / w w w 3 . c m b i . u m c n . n l /…”
Essentiality of LD-Transpeptidation in Agrobacterium tumefaciensPreprint
Aliashkevich, A., Guest, T., Alvarez, L., Gilmore, M. C., Amstutz, J., Mateus, A., Schiffthaler, B., Ruiz, I., Typas, A., Savitski, M. M., et al. (2024) · bioRxiv (Cold Spring Harbor Laboratory) · doi:10.1101/2024.06.21.600065 · Cited by 1
Woodard, J., Liu, Z., Chegini, A. M., Tian, J., Bhowmick, R., Pennathur, S., Mashaghi, A., Brender, J. R., & Chandrasekaran, S. (2025) · Cell Reports Methods · doi:10.1016/j.crmeth.2025.101169 · Cited by 3
Metadata
- Authors
- Varadi, M., Bertoni, D., Magana, P., Paramval, U., Pidruchna, I., Radhakrishnan, M., Tsenkov, M., Nair, S., Mirdita, M., Yeo, J., Kovalevskiy, O., Tunyasuvunakool, K., Laydon, A., Žídek, A., Tomlinson, H., Hariharan, D., Abrahamson, J., Green, T., Jumper, J., Birney, E., Steinegger, M., Hassabis, D., & Velankar, S.
- Affiliations
- European Molecular Biology Laboratory, European Bioinformatics Institute , Hinxton, UK
- European Bioinformatics Institute (GB)
- Seoul National University (KR)
- Google DeepMind (United Kingdom) (GB)
- Corresponding author
- Hassabis, Demis
- Velankar, Sameer
- ORCID
- Venue
- Nucleic Acids Research
- Publisher
- Oxford University Press (OUP)
- Published
- 2023-11-02
- Volume
- 52
- Issue
- D1
- Pages
- D368-D375
- ISSN
- 0305-1048 (ISSN-L)
- 1362-4962
- PMC
- PMC10767828
- OpenAlex
- W4388464011
- Cited by
- 2,052
- References
- 29
- Citations by year
- 2026: 724 · 2025: 1015 · 2024: 307 · 2023: 5
- FWCI
- 181.98
- Type
- journal article
- Language
- en
- Open access
- gold
- doi.org
- Licence
- Locations
- Nucleic Acids Research (publishedVersion, cc-by)
- PubMed Central (publishedVersion, cc-by)
- PubMed (publishedVersion)
- Seoul National University Open Repository (Seoul National University) (submittedVersion)
- Funders
- Google DeepMind
- National Research Foundation of Korea (2019R1A6A1A10073437, 2020M3A9G7103933, 2021R1C1C102065, 2021M3A9I4021220, RS-2023-00250470)
- Samsung DS Research Fund
- Seoul National University
- Topics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Protein Structure and Dynamics
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Structural Biology › Enzyme Structure and Function
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Molecular Biology › Machine Learning in Bioinformatics
- Keywords
- predicted protein structures, AlphaFold2, protein structure prediction, protein sequence coverage, data archiving, database access, search engines
- MeSH terms
- Amino Acid Sequence, Artificial Intelligence, Proteins, Protein Structure, Secondary, Proteome, Databases, Protein, Search Engine
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
The claim verdicts this report counts are experimental: their accuracy is not yet measured.
A language model drew the claims from the paper on 2026-10-10, under version 3 of the claim definition (model deepseek-v4.1-flash, prompt extract-v3).
A second model labelled each claim on 2026-10-10 (model jev-1.13-20260917, definition revision gold-guideline-v0.2).
Cited sources are checked with a language model’s help (gpt-6-sol).
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
Data sources
Bibliographic data: Semantic Scholar, OpenAlex, Crossref.
Contains information from the Semantic Scholar Academic Graph Datasets, made available under the ODC Attribution License (ODC-BY 1.0).
Semantic Scholar’s data is described in: Kinney, R., et al. (2023). The Semantic Scholar Open Data Platform. arXiv:2301.10140.
