

The DOI matches its Crossref record; 382 of the 390 references it cites were found in a registry or an index; 220 of the 329 sources its claims cite were checked for support.
IL-6 in inflammation, autoimmunity and cancer
Open accessAbstract
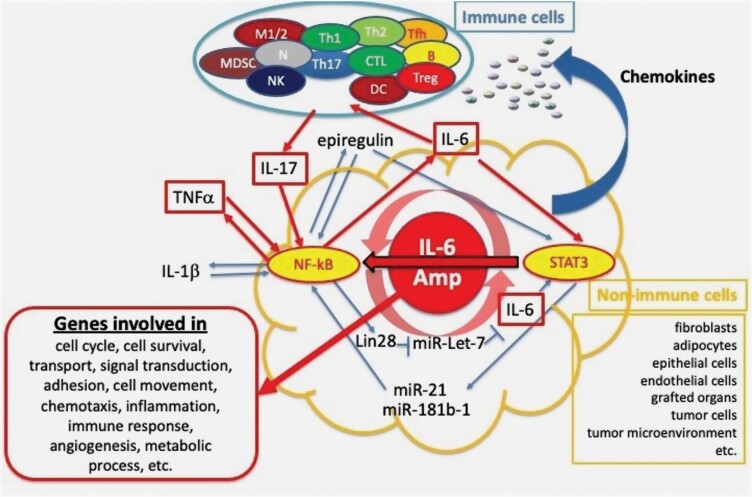
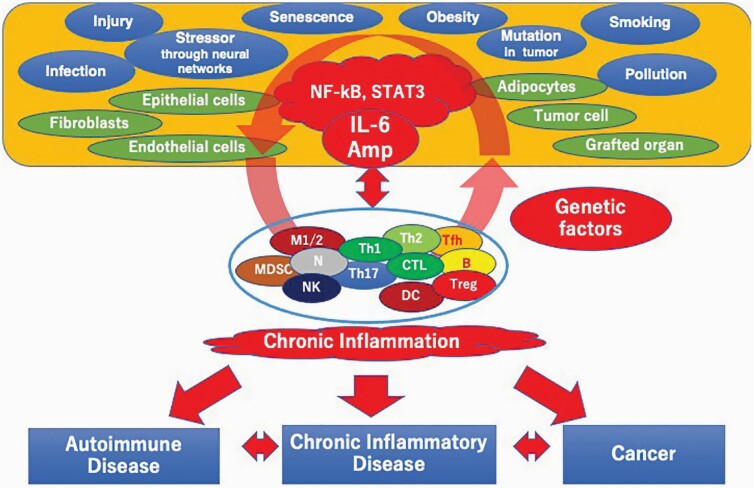
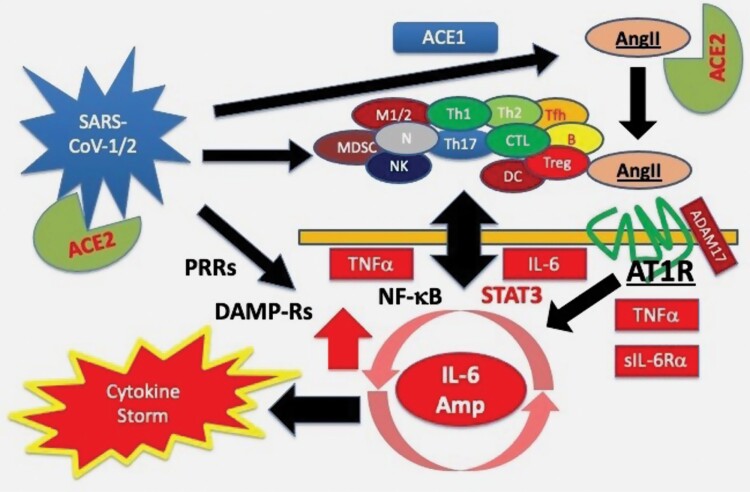
IL-6 is involved both in immune responses and in inflammation, hematopoiesis, bone metabolism and embryonic development. IL-6 plays roles in chronic inflammation (closely related to chronic inflammatory diseases, autoimmune diseases and cancer) and even in the cytokine storm of corona virus disease 2019 (COVID-19). Acute inflammation during the immune response and wound healing is a well-controlled response, whereas chronic inflammation and the cytokine storm are uncontrolled inflammatory responses. Non-immune and immune cells, cytokines such as IL-1β, IL-6 and tumor necrosis factor alpha (TNFα) and transcription factors nuclear factor-kappa B (NF-κB) and signal transducer and activator of transcription 3 (STAT3) play central roles in inflammation. Synergistic interactions between NF-κB and STAT3 induce the hyper-activation of NF-κB followed by the production of various inflammatory cytokines. Because IL-6 is an NF-κB target, simultaneous activation of NF-κB and STAT3 in non-immune cells triggers a positive feedback loop of NF-κB activation by the IL-6–STAT3 axis. This positive feedback loop is called the IL-6 amplifier (IL-6 Amp) and is a key player in the local initiation model, which states that local initiators, such as senescence, obesity, stressors, infection, injury and smoking, trigger diseases by promoting interactions between non-immune cells and immune cells. This model counters dogma that holds that autoimmunity and oncogenesis are triggered by the breakdown of tissue-specific immune tolerance and oncogenic mutations, respectively. The IL-6 Amp is activated by a variety of local initiators, demonstrating that the IL-6–STAT3 axis is a critical target for treating diseases.
Overview
AI snapshot
Written from the paper’s full text.
TL;DR
IL-6 is a pleiotropic cytokine discovered in 1986 that acts through IL-6–STAT3 signaling and the IL-6 amplifier (IL-6 Amp), a STAT3/NF-κB-driven amplification loop, to link inflammation, autoimmunity and cancer. The review proposes a local initiation model in which infection, injury, stress, senescence, obesity and oncogenic mutations activate the IL-6 Amp in non-immune cells, recruiting immune cells and driving chronic disease.
Key findings
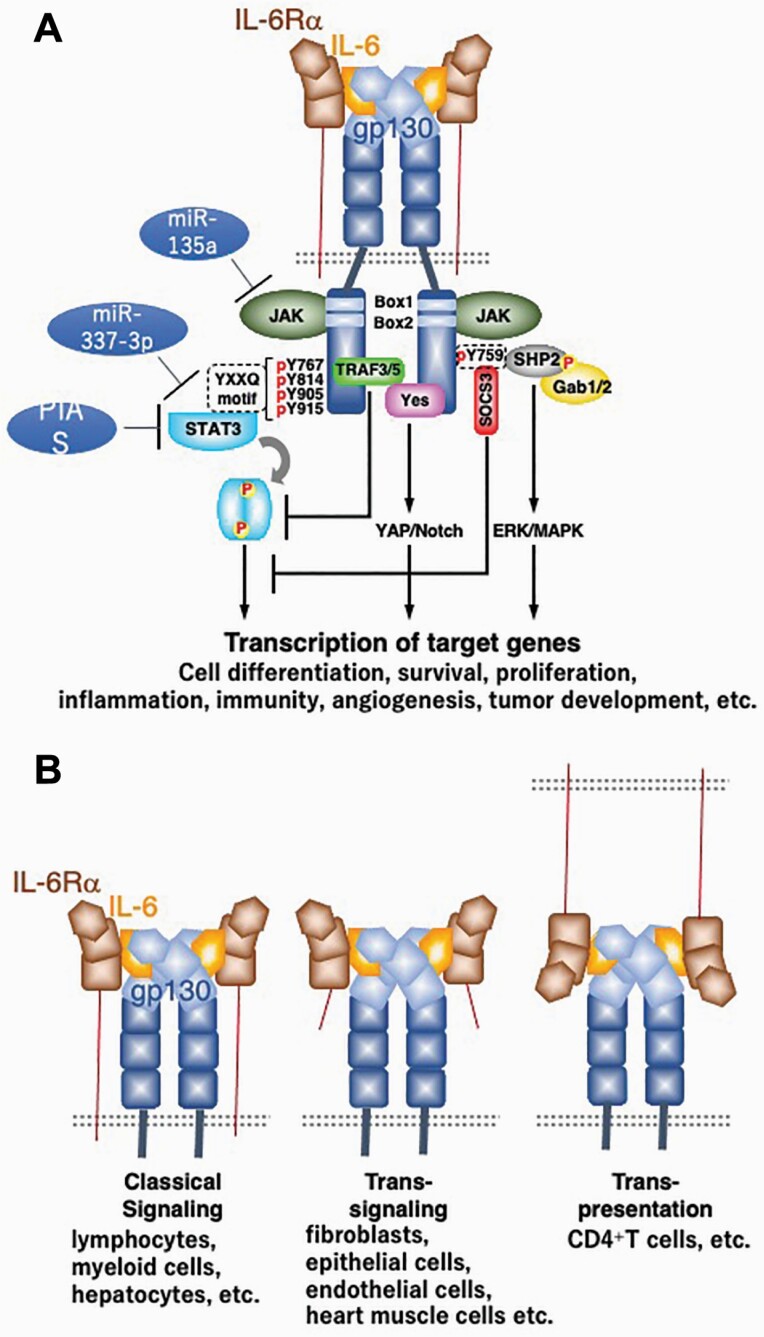
- IL-6 is a prototypical member of the 10-member IL-6 family and signals via classic, trans-signaling and trans-presentation modes; trans-signaling expands IL-6 targets to cells expressing only gp130, including fibroblasts and endothelial cells.
- IL-6–STAT3 signaling regulates Th17/Treg balance, plasma cell differentiation, dendritic cell maturation, M2 macrophages, MDSCs and PD-L1 expression, and promotes cancer cell survival, proliferation, EMT, angiogenesis and metastasis.
- The IL-6 Amp synergistically amplifies IL-6 and other pro-inflammatory mediators through STAT3 and NF-κB; a comparative analysis found 202 of 1700 screened genes showed over 490 indications of association with autoimmune, metabolic, neurodegenerative and other chronic inflammatory diseases.
- IL-6 correlates with prognosis across 23 cancer types; in mouse models, F759 mice spontaneously develop IL-6-dependent, CD4+ T-cell-dependent arthritis, while gp130Y757F/Y757F mice show gastric hyperproliferative lesions as early as 6 weeks.
Methods
- Narrative review synthesizing mechanistic, genetic, mouse-model and clinical literature on IL-6 in inflammation, autoimmunity and cancer.
- Discusses mouse models including F759, gp130Y757F/Y757F, IL-6 transgenic and IL-6-deficient mice, plus GWAS/SNP associations and clinical trials of anti-IL-6/IL-6R therapies.
- No systematic search strategy, meta-analysis or primary data collection is described.
Limitations
- Stated: many questions remain, including how sIL-6Rα is generated, the mechanism of STAT3–NF-κB synergy, and the genetic/environmental factors that destabilize the IL-6 Amp; current knowledge is insufficient to predict disease end stage or detailed process.
- Stated: more study is needed on interactions among pro-inflammatory molecules and on real-time dynamics of receptor complexes and signaling molecules.
- Evident from the text: as a narrative review, it does not systematically assess study quality or bias, and many mechanistic conclusions derive from mouse models and in vitro systems rather than human validation.
Model-written summary. Check it against the paper before relying on it.
Integrity signals
Registry
Cites
Statistics
Figures
Verification
Claims the paper makes
71 Partly supported · 43 Supported · 41 Own interpretation · 31 No full text of the sources · 10 Supported so far · 7 No basis given · 6 Couldn’t confirm · 3 Not checked yet · accuracy not yet measured
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Consistently, evidence has indicated that interactions between immune cells and non-immune tissues play critical roles in chronic inflammatory diseases and autoimmune diseases (3, 41, 49–51).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [3] Ishihara, 2002, IL-6 in autoimmune disease and chronic inflammatory proliferative disease, Cytokine Growth Factor Rev. source
- Partly supportedcites [41] Hirano, 2010, Interleukin 6 in autoimmune and inflammatory diseases: a personal memoir, Proc. Jpn Acad. Ser. B. Phys. Biol. Sci. sourceExperimental — accuracy not yet measured
In “The view from the mountaintop,” the paper says its model implies that interactions between nonimmune and immune systems may play critical roles in autoimmune diseases and chronic inflammatory proliferative disease. The citing sentence presents that tentative, model-based conclusion as an established role across chronic inflammatory diseases more broadly. (LLM, conf=0.84)
- Not checked — no full text of the sourcecites [49] Matsumoto, 2002, How antibodies to a ubiquitous cytoplasmic enzyme may provoke joint-specific autoimmune disease, Nat. Immunol. source
- Not checked — no full text of the sourcecites [50] Marrack, 2001, Autoimmune disease: why and where it occurs, Nat. Med. source
- Not checked — no full text of the sourcecites [51] Hirano, 2002, Revival of the autoantibody model in rheumatoid arthritis, Nat. Immunol. source
hedgedcorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
The sources with a verdict support this claim; the rest weren’t checked yet, have no full text we could read, or their verdicts aren’t rated yet.
In the paper
“Furthermore, tissue property-dependent events (49), microbleeding (52, 53), neural stimulation (54, 55), cell senescence (56, 57) and oncogenic mutations in pre-neoplastic cells have been shown to trigger these interactions.”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [49] Matsumoto, 2002, How antibodies to a ubiquitous cytoplasmic enzyme may provoke joint-specific autoimmune disease, Nat. Immunol. source
- Supportedcites [52] Murakami, 2011, Local microbleeding facilitates IL-6- and IL-17-dependent arthritis in the absence of tissue antigen recognition by activated T cells, J. Exp. Med. source
This citation is attached to microbleeding, not to the other events listed in the sentence. The Discussion describes microbleeding as a local event that induces tissue-specific accumulation of activated T cells, which can then enhance IL-6 signaling in the affected tissue. (LLM, conf=0.90)
- Not checked — no full text of the sourcecites [53] Leavy, 2011, Autoimmunity: joint damage without antigen, Nat. Rev. Immunol. source
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
The sources with a verdict support this claim; the rest weren’t checked yet, have no full text we could read, or their verdicts aren’t rated yet.
In the paper
“Additionally, interactions between a tumor and its microenvironment through the immune system are critical for tumor development and progression (2, 6, 9, 58–60)”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [2] Balkwill, 2001, Inflammation and cancer: back to Virchow?, Lancet source
- Supportedcites [6] Mantovani, 2008, Cancer-related inflammation, Nature source
The opening section (p. 1) says inflammation in the tumour microenvironment promotes tumour development, survival, angiogenesis and metastasis. The adaptive-immunity section (p. 5) and Figure 4 (p. 6) describe immune cells and signalling within that environment affecting tumour development and progression. (LLM, conf=0.93)
- Supportedcites [9] Grivennikov, 2010, Immunity, inflammation, and cancer, Cell source
The “Immune cells in tumorigenesis” section says that immune cells and cancer cells in the tumor microenvironment communicate to control and shape tumor growth. It also explains that immune mediators can either inhibit tumor development or enhance progression, supporting the claim without implying that every immune interaction promotes cancer. (LLM, conf=0.93)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“It too plays a key role in inflammatory diseases including cytokine storm syndromes, autoimmune diseases and cancer (10, 11, 15, 41, 66–68)”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Partly supportedcites [10] Murakami, 2013, Disease-association analysis of an inflammation-related feedback loop, Cell Rep. sourceExperimental — accuracy not yet measured
The Discussion (page not specified) describes the IL-6 amplifier as important in chronic inflammation and reports associations with autoimmune diseases, including rheumatoid arthritis and lupus. The passages found do not support the added claims about cytokine storm syndromes or cancer. (LLM, conf=0.88)
- Not checked — no full text of the sourcecites [11] Atsumi, 2014, Inflammation amplifier, a new paradigm in cancer biology, Cancer Res. source
- Partly supportedcites [15] Hirano, 2020, COVID-19: a new virus, but a familiar receptor and cytokine release syndrome, Immunity sourceExperimental — accuracy not yet measured
The Main Text says the IL-6 amplifier can lead to inflammatory and autoimmune diseases. It also proposes a possible role in cytokine release syndrome, rather than establishing a key role. Cancer is mentioned as the condition treated by CAR-T cells, not as a disease in which the IL-6 amplifier plays a key role. (LLM, conf=0.87)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“The IL-6–STAT3 pathway is now considered a major target for the treatment of inflammatory diseases (15, 19, 21, 24–26, 76).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Partly supportedcites [15] Hirano, 2020, COVID-19: a new virus, but a familiar receptor and cytokine release syndrome, Immunity sourceExperimental — accuracy not yet measured
The Main Text (page unavailable) identifies the IL-6–STAT3 axis as a potential target for cytokine release syndrome in later-stage COVID-19 and links IL-6–STAT3 signaling to inflammatory and autoimmune diseases. It does not establish the pathway as a major treatment target for inflammatory diseases generally; that broader, more definite claim exceeds the paper's qualified proposal. (LLM, conf=0.86)
- Partly supportedcites [19] Yu, 2009, STATs in cancer inflammation and immunity: a leading role for STAT3, Nat. Rev. Cancer sourceExperimental — accuracy not yet measured
In “Targeting STAT3 for cancer therapy” (page not supplied), the paper calls JAK kinases in IL-6–STAT3 signalling crucial targets for inflammation-induced cancer and cancer-associated inflammation. That supports targeting the pathway in a cancer-related setting, but not the broader claim that it is a major treatment target for inflammatory diseases generally. (LLM, conf=0.86)
- Not checked — no full text of the sourcecites [21] Hunter, 2015, IL-6 as a keystone cytokine in health and disease, Nat. Immunol. source
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“However, this site is also a recruitment motif for SOCS3, and it was found that the F759/F757-dependent negative regulation is mainly dependent on the recruitment of SOCS3 (116, 117).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Partly supportedcites [116] Lehmann, 2003, SHP2 and SOCS3 contribute to Tyr-759-dependent attenuation of interleukin-6 signaling through gp130, J. Biol. Chem. sourceExperimental — accuracy not yet measured
The opening summary and Results report that SOCS3 acts through gp130 Tyr-759 and that its inhibitory activity depends on that site. But the paper also finds an independent inhibitory role for SHP2 and describes two largely distinct modes of negative regulation; it does not establish that SOCS3 recruitment accounts for most of the site-dependent effect. Our copy addresses Tyr-759, not the cited F757 counterpart. (LLM, conf=0.84)
- Supportedcites [117] Fairlie, 2003, Negative regulation of gp130 signalling mediated through tyrosine-757 is not dependent on the recruitment of SHP2, Biochem. J. source
The Introduction reports that a gp130 receptor mutant with reduced SOCS3 recruitment but retained SHP2 binding showed elevated cytokine responsiveness, like a Y757F mutant. The Discussion identifies Y757 as the shared docking site. Together, these passages support the claim that Y757-dependent negative regulation depends mainly on SOCS3 recruitment; the paper does not separately test F759. (LLM, conf=0.90)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
The cited sources that could be checked couldn’t confirm or dispute this claim.
In the paper
“Indeed, this mode plays crucial roles in chronic inflammation (33, 96).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Could not verifycites [33] Murakami, 2019, Pleiotropy and specificity: insights from the interleukin 6 family of cytokines, Immunity sourceExperimental — accuracy not yet measured
The supplied passage has no section or page label and consists of reference links, not discussion of trans-signaling. Searches of our copy for “trans-signaling,” “chronic inflammation,” and related IL-6 signaling terms found no substantive passage addressing the claim. (LLM, conf=0.72)
- Not checked — no full text of the sourcecites [96] Rose-John, 2017, The soluble interleukin 6 receptor: advanced therapeutic options in inflammation, Clin. Pharmacol. Ther. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Could not verify
Every source the paper cites for this claim was checked and supports it.
In the paper
“STAT3-induced microRNA (miR)-21 and miR-181b-1 activate NF-κB, resulting in the induction of IL-6 gene expression (90).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Supportedcites [90] Iliopoulos, 2010, STAT3 activation of miR-21 and miR-181b-1 via PTEN and CYLD are part of the epigenetic switch linking inflammation to cancer, Mol. Cell source
The Results section reports that STAT3 activates miR-21 and miR-181b-1. Its CYLD subsection links miR-181b-1 to increased NF-κB activity and IL6 production; its PTEN/Akt subsection links miR-21 to NF-κB activation and reports that Akt inhibition reduces IL6 production. Page numbers are not provided in our copy. (LLM, conf=0.91)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Supported
The cited sources that could be checked couldn’t confirm or dispute this claim.
In the paper
“The IL-6R is composed of an IL-6-binding receptor molecule (IL-6Rα) and a signal transducer, gp130, which is shared among the IL-6 family of cytokines (Fig. 1A) (33).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Could not verifycites [33] Murakami, 2019, Pleiotropy and specificity: insights from the interleukin 6 family of cytokines, Immunity sourceExperimental — accuracy not yet measured
The passages from our copy contain reference links and corrupted section headings, but no readable discussion of IL-6R composition. Searches for IL-6Rα, gp130, and shared IL-6-family receptor subunits returned no passage that addresses the claim. (LLM, conf=0.72)
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Could not verify
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Similarly, defective STAT3 activation enhances SHP-2–ERK–MAPK activation pathways, most likely through an impaired STAT3-mediated induction of suppressor of cytokine signaling 3 (SOCS3) (111, 112).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Partly supportedcites [111] Ernst, 2001, Defective gp130-mediated signal transducer and activator of transcription (STAT) signaling results in degenerative joint disease, gastrointestinal ulceration, and failure of uterine implantation, J. Exp. Med. sourceExperimental — accuracy not yet measured
The Results section, “Sustained Activation of the SHP-2/ras/Erk Pathway Results from Impaired Induction of SOCS-1” (page unavailable), links impaired STAT signaling to sustained SHP-2/ras/Erk activation and reports reduced SOCS-3 induction. It does not establish the specific STAT3-to-SOCS3 mechanism claimed: it emphasizes SOCS-1 and describes induction of both SOCS-1 and SOCS-3 as mediated at least in part by STATs. (LLM, conf=0.86)
- Supportedcites [112] Jenkins, 2004, Imbalanced gp130-dependent signaling in macrophages alters macrophage colony-stimulating factor responsiveness via regulation of c-fms expression, Mol. Cell. Biol. source
The Discussion reports that loss of gp130-dependent STAT1/3 signaling increases IL-6-induced SHP2 and ERK1/2 phosphorylation because STAT-mediated induction of SOCS1 and SOCS3 is impaired. The Results also show enhanced ERK1/2 phosphorylation after IL-6 stimulation in macrophages with defective gp130-dependent STAT signaling. The claim captures this mechanism, although the paper names SOCS1 alongside SOCS3. (LLM, conf=0.90)
hedgedconditional- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
- Partly supported
- It may state two separate claims as one. 78
- Probably not a substantial claim. 7
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Among them, IL-6 is a major player in chronic inflammatory diseases, autoimmune diseases, cancer and cytokine storm (1, 3, 10–26).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [1] Hirano, 1998, Interleukin 6 and its receptor: ten years later, Int. Rev. Immunol. source
- Not checked — no full text of the sourcecites [3] Ishihara, 2002, IL-6 in autoimmune disease and chronic inflammatory proliferative disease, Cytokine Growth Factor Rev. source
- Partly supportedcites [10] Murakami, 2013, Disease-association analysis of an inflammation-related feedback loop, Cell Rep. sourceExperimental — accuracy not yet measured
The Introduction describes IL-6 as important in autoimmune diseases and other inflammatory conditions, and the Discussion links the IL-6 amplifier to chronic inflammation in humans. The passages do not support the additional claims about cancer or cytokine storm; searches for those terms found no relevant discussion in our copy. (LLM, conf=0.86)
- Not checked — no full text of the sourcecites [11] Atsumi, 2014, Inflammation amplifier, a new paradigm in cancer biology, Cancer Res. source
- Partly supportedcites [12] Mehta, 2020, COVID-19: consider cytokine storm syndromes and immunosuppression, Lancet sourceExperimental — accuracy not yet measured
The main text links elevated IL-6 to fatal COVID-19 cases and discusses IL-6 receptor blockade for cytokine release syndrome, supporting its relevance to cytokine storm. It does not address IL-6's role in chronic inflammatory diseases, autoimmune diseases, or cancer. The supplied copy has no page numbers. (LLM, conf=0.87)
- Partly supportedcites [13] Mahmudpour, 2020, COVID-19 cytokine storm: the anger of inflammation, Cytokine sourceExperimental — accuracy not yet measured
The ACE/Angiotensin II/AT1R axis section describes IL-6 signaling as part of a mechanism that induces cytokine storm in severe COVID-19. The Introduction also reports higher IL-6 levels in patients who died of COVID-19 than in those who recovered. These passages support the cytokine-storm part of the claim, but the paper does not establish IL-6 as a major player in chronic inflammatory diseases, autoimmune diseases, or cancer. (LLM, conf=0.87)
- Not checked — no full text of the sourcecites [14] Hu, 2021, The cytokine storm and COVID-19, J. Med. Virol. source
- Partly supportedcites [15] Hirano, 2020, COVID-19: a new virus, but a familiar receptor and cytokine release syndrome, Immunity sourceExperimental — accuracy not yet measured
The Main Text describes the IL-6 amplifier as contributing to inflammatory and autoimmune diseases and discusses IL-6 signaling in COVID-19 cytokine release syndrome. It does not establish IL-6 as a major player in cancer generally; its mention of leukemia and lymphoma concerns cytokine release syndrome following CAR-T therapy. Page numbers are unavailable in our copy. (LLM, conf=0.85)
- Partly supportedcites [16] García, 2020, Immune response, inflammation, and the clinical spectrum of COVID-19, Front. Immunol. sourceExperimental — accuracy not yet measured
The Immunological Profile of COVID-19 Patients (page not provided) identifies IL-6 among elevated inflammatory markers in severe COVID-19 and says this inflammatory response may result in cytokine storm. The Changes in Cytokine/Chemokine Plasma Levels section also links higher IL-6 levels to more severe COVID-19. These passages do not establish IL-6 as a major player in chronic inflammatory diseases, autoimmune diseases, or cancer. (LLM, conf=0.87)
- Partly supportedcites [17] Vabret, 2020, Immunology of COVID-19: current state of the science, Immunity sourceExperimental — accuracy not yet measured
The Therapeutic Immunomodulation for COVID-19 Treatment section (page unavailable) describes IL-6 as a critical cytokine in COVID-19-associated cytokine release syndrome, supporting the cytokine-storm portion of the claim. The passages found do not establish that IL-6 is a major player in chronic inflammatory diseases, autoimmune diseases, or cancer. (LLM, conf=0.84)
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The cited sources that could be checked couldn’t confirm or dispute this claim.
In the paper
“IL-6 is a prototypical member of the IL-6 family of cytokines, which is composed of 10 members including IL-6, IL-11, IL-27, oncostatin M (OSM), leukemia inhibitory factor (LIF), ciliary neurotrophic factor (CNTF), cardiotrophin 1 (CT-1), cardiotrophin-like cytokine factor 1 (CLCF1), IL-35 and IL-39 (33).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Could not verifycites [33] Murakami, 2019, Pleiotropy and specificity: insights from the interleukin 6 family of cytokines, Immunity sourceExperimental — accuracy not yet measured
The supplied passages have no section or page labels and contain only reference links, not discussion of the IL-6 family. Searches for the ten-member list, IL-39, and “prototypical member” returned no substantive passage, so our copy cannot verify the claimed membership or count. (LLM, conf=0.72)
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it. It may state two separate claims as one.
- Could not verify
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“IL-6, which was discovered in 1986 (27), is a pleiotropic cytokine involved not only in immune responses but also in inflammation, hematopoiesis, bone metabolism, embryonic development and other fundamental processes (1, 21, 28–32).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [1] Hirano, 1998, Interleukin 6 and its receptor: ten years later, Int. Rev. Immunol. source
- Not checked — no full text of the sourcecites [21] Hunter, 2015, IL-6 as a keystone cytokine in health and disease, Nat. Immunol. source
- Not checked — no full text of the sourcecites [28] Van Snick, 1990, Interleukin-6: an overview, Annu. Rev. Immunol. source
- Not checked — no full text of the sourcecites [29] Hirano, 1990, Interleukin-6. source
- Partly supportedcites [30] Heinrich, 2003, Principles of interleukin (IL)-6-type cytokine signalling and its regulation, Biochem. J. sourceExperimental — accuracy not yet measured
The Abstract and Introduction describe IL-6-type cytokines, a family that includes IL-6, as involved in immune responses, inflammation, haematopoiesis and embryonal development. The Abstract also links dysregulated signalling in this family to osteoporosis, but it does not specifically state that IL-6 itself is involved in bone metabolism or each of the other listed processes. (LLM, conf=0.83)
- Not checked — no full text of the sourcecites [31] Kamimura, 2003, IL-6 signal transduction and its physiological roles: the signal orchestration model, Rev. Physiol. Biochem. Pharmacol. source
- Partly supportedcites [32] Hasegawa, 2016, Expanding diversity in molecular structures and functions of the IL-6/IL-12 heterodimeric cytokine family, Front. Immunol. sourceExperimental — accuracy not yet measured
The Introduction describes IL-6 as an important inflammatory cytokine, and the Abstract describes pleiotropic immune functions of the IL-6/IL-12 family. The copy does not establish the claimed roles of IL-6 specifically in hematopoiesis, bone metabolism, or embryonic development; its hematopoiesis discussion concerns IL-27. (LLM, conf=0.86)
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“IL-6 is also a key factor in inflammation, autoimmunity and cancer, with its effect mainly exerted through the IL-6–signal transducer and activator of transcription 3 (STAT3) pathway (18–26, 61–65).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Partly supportedcites [18] Grivennikov, 2009, IL-6 and Stat3 are required for survival of intestinal epithelial cells and development of colitis-associated cancer, Cancer Cell sourceExperimental — accuracy not yet measured
The Introduction describes IL-6 as important in chronic inflammation, T-cell-dependent autoimmune disorders and cancer. The Discussion identifies STAT3 as a major IL-6 effector in colitis-associated cancer, but also lists two other IL-6 signaling pathways. It does not establish that IL-6 acts mainly through STAT3 across all three settings. (LLM, conf=0.85)
- Partly supportedcites [19] Yu, 2009, STATs in cancer inflammation and immunity: a leading role for STAT3, Nat. Rev. Cancer sourceExperimental — accuracy not yet measured
The ‘STAT proteins in immune modulation’ section describes IL-6 signalling through GP130–JAK to activate STAT3, and ‘STAT3 induces cancer inflammation’ describes IL-6-driven STAT3 activation in myeloma. The paper also calls STAT3 the major IL-6R-dependent transcription factor in T cells (‘STATs in anti-cancer or pro-cancer inflammation’). These passages support the inflammation-and-cancer link, but do not establish that IL-6 acts mainly through STAT3 across all the claimed contexts or directly characterize IL-6 as a key factor in autoimmunity; autoimmunity appears in reference titles. (LLM, conf=0.83)
- Partly supportedcites [20] Bollrath, 2009, gp130-mediated Stat3 activation in enterocytes regulates cell survival and cell-cycle progression during colitis-associated tumorigenesis, Cancer Cell sourceExperimental — accuracy not yet measured
The Discussion (page unavailable) reports that IL-6-dependent STAT3 activation promotes colitis-associated cancer, supporting the inflammation-and-cancer portion of the claim. Our copy does not address autoimmunity or establish that IL-6 acts mainly through STAT3 across all three conditions; the Significance section also describes other downstream gp130 pathways. (LLM, conf=0.87)
- Not checked — no full text of the sourcecites [21] Hunter, 2015, IL-6 as a keystone cytokine in health and disease, Nat. Immunol. source
- Partly supportedcites [22] Masjedi, 2018, The significant role of interleukin-6 and its signaling pathway in the immunopathogenesis and treatment of breast cancer, Biomed. Pharmacother. sourceExperimental — accuracy not yet measured
Section 2 links IL-6 to inflammation, and section 6.2 discusses IL-6 blockade in autoimmune disorders. Sections 3 and 5 describe IL-6/JAK/STAT3 signaling and call STAT3 the most important player in IL-6’s cancer-related functions. The paper does not establish that IL-6’s effects in inflammation and autoimmunity are mainly exerted through STAT3. (LLM, conf=0.82)
- Partly supportedcites [23] Gyamfi, 2018, Interleukin-6/STAT3 signalling regulates adipocyte induced epithelial-mesenchymal transition in breast cancer cells, Sci. Rep. sourceExperimental — accuracy not yet measured
The Introduction and Discussion link IL-6 to breast cancer progression through STAT3 signalling, and the Results show reduced STAT3 activity when IL-6 signalling was blocked. The cited paper does not address autoimmunity or establish that IL-6 acts mainly through STAT3 across inflammation, autoimmunity and cancer; its findings are specific to adipocyte–breast cancer cell interactions. Page numbers are not available in our copy. (LLM, conf=0.88)
- Supportedcites [24] Jones, 2018, Recent insights into targeting the IL-6 cytokine family in inflammatory diseases and cancer, Nat. Rev. Immunol. source
The ‘Determinants of chronic autoimmune diseases’ section describes IL-6-driven joint inflammation and says STAT3 primarily regulates these disease features. ‘Contrasting roles in cancer’ describes IL-6 as pro-tumourigenic and identifies STAT3-mediated oncogenic processes; Figure 2 also describes predominant STAT3 activation. Together, these passages support the claim’s broad characterization, although the paper discusses other IL-6 signalling routes as well. Page numbers were not supplied in our copy. (LLM, conf=0.89)
- Partly supportedcites [25] Rebe, 2019, STAT3, a master regulator of anti-tumor immune response, Cancer sourceExperimental — accuracy not yet measured
Section 3.2 (Dendritic Cells; page not available) describes IL-6 acting through gp130–STAT3 in immune-cell differentiation, and Section 3.1 (Macrophages) links IL-6/STAT3 signaling to pro-tumor effects. Section 2.2 (Th17) links IL-6-induced cells to autoimmune disorders. These passages support roles for IL-6 in the named contexts, but do not establish that its effects across inflammation, autoimmunity, and cancer are mainly exerted through STAT3. (LLM, conf=0.84)
- Partly supportedcites [26] Johnson, 2018, Targeting the IL-6/JAK/STAT3 signalling axis in cancer, Nat. Rev. Clin. Oncol. sourceExperimental — accuracy not yet measured
The IL-6 section describes IL-6 as a major mediator of inflammation and important in cancer, and discusses arthritis as an inflammatory condition involving IL-6. The STAT3 section says JAK/STAT3 mediates many IL-6 effects in tumours, but it does not establish that IL-6 acts mainly through STAT3 across inflammation, autoimmunity and cancer; the JAKs section also identifies other signalling pathways. (LLM, conf=0.83)
- Not checked — no full text of the sourcecites [61] Bromberg, 2000, The role of STATs in transcriptional control and their impact on cellular function, Oncogene source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Instead of murine MoAb against IL-6, a humanized MoAb against IL-6 receptor (IL-6R), tocilizumab, has proven successful for treating RA and Castleman’s disease (71–75).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [71] Choy, 2002, Therapeutic benefit of blocking interleukin-6 activity with an anti-interleukin-6 receptor monoclonal antibody in rheumatoid arthritis: a randomized, double-blind, placebo-controlled, dose-escalation trial, Arthritis Rheum. source
- Partly supportedcites [72] Nishimoto, 2005, Humanized anti-interleukin-6 receptor antibody treatment of multicentric Castleman disease, Blood sourceExperimental — accuracy not yet measured
The Study drug section identifies MRA as a humanized anti–IL-6 receptor antibody, and the Discussion reports that it alleviated symptoms in patients with multicentric Castleman disease. The paper does not report successful treatment of rheumatoid arthritis. Page numbers are not available in our copy. (LLM, conf=0.88)
- Not checked — no full text of the sourcecites [73] Maini, 2006, Double-blind randomized controlled clinical trial of the interleukin-6 receptor antagonist, tocilizumab, in European patients with rheumatoid arthritis who had an incomplete response to methotrexate, Arthritis Rheum. source
- Not checked — no full text of the sourcecites [74] Genovese, 2008, Interleukin-6 receptor inhibition with tocilizumab reduces disease activity in rheumatoid arthritis with inadequate response to disease-modifying antirheumatic drugs: the tocilizumab in combination with traditional disease-modifying antirheumatic drug therapy study, Arthritis Rheum. source
- Partly supportedcites [75] Jones, 2010, Comparison of tocilizumab monotherapy versus methotrexate monotherapy in patients with moderate to severe rheumatoid arthritis: the AMBITION study, Ann. Rheum. Dis. sourceExperimental — accuracy not yet measured
The paper identifies tocilizumab as a humanised anti-IL-6 receptor antibody (introduction, following the trial registration number). Its Results and Discussion report efficacy for rheumatoid arthritis. Our copy does not address its use for Castleman’s disease, so that part of the claim is unsupported by this source. (LLM, conf=0.90)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Notably, IL-6 is involved in cell senescence, is produced by senescent cells and is critically involved in senescence-induced inflammation and age-dependent pathologies and cancer (7).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Partly supportedcites [7] Kuilman, 2008, Oncogene-induced senescence relayed by an interleukin-dependent inflammatory network, Cell sourceExperimental — accuracy not yet measured
The Results show that IL-6 is produced by senescent cells and that IL-6/IL-6R is essential for oncogene-induced senescence; the paper’s Summary says IL-6 depletion collapses the associated inflammatory network. The paper proposes a connection with cancer, but these findings do not establish that IL-6 is critically involved in age-dependent pathologies or in cancer generally. No passage matched a search for “age-dependent.” (LLM, conf=0.86)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
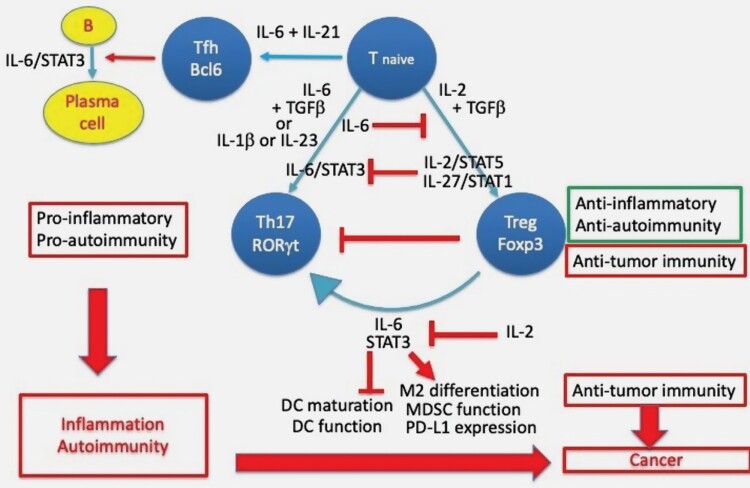
“Indeed, IL-6 promotes antibody production by directly acting on plasma cells and by indirectly promoting Bcl6-dependent follicular CD4(+) T (Tfh) cell differentiation together with IL-21 (Fig. 2) (143–146).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
What the sources say
- Partly supportedcites [143] Muraguchi, 1988, The essential role of B cell stimulatory factor 2 (BSF-2/IL-6) for the terminal differentiation of B cells, J. Exp. Med. sourceExperimental — accuracy not yet measured
The paper’s summary reports that IL-6 (BSF-2) directly acts on PWM-induced B blast cells to induce immunoglobulin production, supporting the direct antibody-production component. It does not establish direct action on plasma cells or address Bcl6-dependent Tfh differentiation or cooperation with IL-21. (LLM, conf=0.87)
- Partly supportedcites [144] Jego, 2003, Plasmacytoid dendritic cells induce plasma cell differentiation through type I interferon and interleukin 6, Immunity sourceExperimental — accuracy not yet measured
The Results (Figure 5) show that IL-6 helps activated B cells become Ig-secreting plasma cells and that blocking IL-6 reduces IgG secretion; in these experiments, IL-6 acts with type I interferon. The paper does not support the attributed mechanism involving Bcl6-dependent Tfh-cell differentiation or IL-21. Page numbers are not available in our copy. (LLM, conf=0.90)
- Partly supportedcites [145] Cassese, 2003, Plasma cell survival is mediated by synergistic effects of cytokines and adhesion-dependent signals, J. Immunol. sourceExperimental — accuracy not yet measured
The Discussion and Figure 4 describe IL-6 acting directly on isolated plasma cells to support their survival and antibody-secreting function in culture. The paper does not address IL-21 or Bcl6-dependent Tfh differentiation, so it does not support the indirect pathway in the citing sentence. It also reports that IL-6 was not essential for maintaining antibody titers in vivo. (LLM, conf=0.88)
- Partly supportedcites [146] Vinuesa, 2016, Follicular helper T cells, Annu. Rev. Immunol. sourceExperimental — accuracy not yet measured
The section “Regulation of Human and Mouse T Follicular Helper Cells by Cytokines” says IL-6 and IL-21 contribute to Tfh differentiation in mice and that IL-6 induces an early wave of BCL6 expression. The passages do not say IL-6 directly acts on plasma cells to promote antibody production; the plasmablast effect described in “T Follicular Helper Effector Molecules” is attributed to IL-21. (LLM, conf=0.87)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“The IL-6–STAT3 pathway is involved in the epithelial–mesenchymal transition (EMT) (131, 132), tumor migration (133), cancer stemness (130, 134, 135) and the induction of hypoxia-inducible factor-1α (HIF-1α) and vascular endothelial growth factor (VEGF), two key players for angiogenesis (136–140).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 signaling in cell survival, proliferation, metastasis and angiogenesis
What the sources say
- Supportedcites [131] Sullivan, 2009, Interleukin-6 induces an epithelial-mesenchymal transition phenotype in human breast cancer cells, Oncogene source
This source is cited for the EMT portion of the sentence; the migration, stemness and angiogenesis clauses have separate citations. The Introduction describes IL-6 signaling through JAK/STAT3, and the Discussion proposes STAT3 as a mediator between IL-6 and an EMT phenotype in human breast cancer cells. (LLM, conf=0.91)
- Partly supportedcites [132] Yadav, 2011, IL-6 promotes head and neck tumor metastasis by inducing epithelial-mesenchymal transition via the JAK-STAT3-SNAIL signaling pathway, Mol. Cancer Res. sourceExperimental — accuracy not yet measured
The Results sections on IL-6-mediated EMT and STAT3 knockdown support the pathway’s involvement in EMT and cell migration. Our copy does not establish cancer stemness or induction of HIF-1α or VEGF by this pathway; the Discussion mentions VEGF only among cytokines previously associated with poor prognosis. (LLM, conf=0.86)
- Not checked — no full text of the sourcecites [133] Barbieri, 2010, Constitutively active Stat3 enhances neu-mediated migration and metastasis in mammary tumors via upregulation of Cten, Cancer Res. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The cited sources that could be checked couldn’t confirm or dispute this claim.
In the paper
“For IL-6 activity, the expression of both IL-6Rα and gp130 on the cell surface is required, which limits the number of target cells on which IL-6 acts to lymphocytes, myeloid cells and hepatocytes (33).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Could not verifycites [33] Murakami, 2019, Pleiotropy and specificity: insights from the interleukin 6 family of cytokines, Immunity sourceExperimental — accuracy not yet measured
The supplied passages and searches for IL-6Rα/gp130 expression and classic IL-6 signaling returned only reference links, with no substantive text or section/page labels. Our copy therefore provides no passage establishing whether cell-surface expression of both receptors is required or whether IL-6 target cells are limited to the three named groups. (LLM, conf=0.75)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Could not verify
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Notably, sIL-6Rα is increased in several diseases, indicating that trans-signaling plays important roles in the progression of inflammation in many diseases including severe COVID-19 (15, 96).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Partly supportedcites [15] Hirano, 2020, COVID-19: a new virus, but a familiar receptor and cytokine release syndrome, Immunity sourceExperimental — accuracy not yet measured
The Main Text describes formation of sIL-6Rα and IL-6–sIL-6Rα signaling as part of an inflammatory pathway potentially relevant to COVID-19. It does not report that sIL-6Rα is increased in several diseases, and it presents the pathway’s role in COVID-19 as a possibility rather than an established finding. No page number is available in our copy. (LLM, conf=0.85)
- Not checked — no full text of the sourcecites [96] Rose-John, 2017, The soluble interleukin 6 receptor: advanced therapeutic options in inflammation, Clin. Pharmacol. Ther. source
hedgedcorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“Regnase-1 degrades IL-6 mRNA (87), while Arid5a protects IL-6 mRNA from Regnase-1-mediated degradation (88).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Supportedcites [87] Maeda, 2017, Regulation of mRNA stability by CCCH-type zinc-finger proteins in immune cells, Int. Immunol. source
The citation to this paper is attached to the first clause, about Regnase-1 degrading IL-6 mRNA. The Regnase-1 section states that Regnase-1 binds IL-6 mRNA and directly degrades it; the Arid5a clause has a separate citation. (LLM, conf=0.93)
- Supportedcites [88] Masuda, 2013, Arid5a controls IL-6 mRNA stability, which contributes to elevation of IL-6 level in vivo, Proc. Natl Acad. Sci. USA source
In “Arid5a Abrogates the Destabilizing Effect of Regnase-1 on IL-6 mRNA” (Fig. 5B; page not specified), cooverexpression of Arid5a and Regnase-1 increased IL-6 mRNA stability compared with Regnase-1 alone. The authors conclude that Arid5a protects against Regnase-1’s destabilizing effect, directly supporting the part attributed to citation 88. (LLM, conf=0.94)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“IL-6 transcription is induced by a variety of stimuli, such as Toll-like receptor (TLR) ligands, IL-1, TNFα, reactive oxygen species (ROS) and zinc (Zn) and also depends on epigenetic regulation (33, 82, 84–86).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Could not verifycites [33] Murakami, 2019, Pleiotropy and specificity: insights from the interleukin 6 family of cytokines, Immunity sourceExperimental — accuracy not yet measured
The available copy shows a corrupted table of contents with a heading for “Regulation of IL-6 Family Cytokine Expression,” but no readable passage from that section (page unavailable). Searches for Toll-like receptor ligands, zinc, reactive oxygen species, and epigenetic regulation returned only reference links or corrupted headings, so the copy cannot verify the claimed transcriptional regulators. (LLM, conf=0.72)
- Partly supportedcites [82] Yoon, 2010, STAT3 transcriptional factor activated by reactive oxygen species induces IL6 in starvation-induced autophagy of cancer cells, Autophagy sourceExperimental — accuracy not yet measured
The section on STAT3 activation in starvation-induced autophagic cancer cells reports that ROS generated during starvation activates a pathway leading to IL6 induction; its promoter experiments also address IL6 transcription. Our copy does not support the other listed stimuli—TLR ligands, IL-1, TNFα and zinc—or the claim about epigenetic regulation. The paper’s finding is specifically in the context of starved cancer cells. (LLM, conf=0.87)
- Not checked — no full text of the sourcecites [84] Wolf, 2014, Interleukin-6 and its receptors: a highly regulated and dynamic system, Cytokine source
- Supportedcites [85] Nishida, 2019, Mast cells play role in wound healing through the ZnT2/GPR39/IL-6 axis, Sci. Rep. source
The Results section, “Zn can induce Il-6 gene expression through a Zn receptor GPR39” (page not available), reports that zinc induced Il-6 mRNA in the tested cell types. The Results section on the PKC/MAPK/C/EBP pathway also links zinc stimulation to Il-6 promoter activity, supporting the zinc portion of this multiply cited sentence. (LLM, conf=0.90)
- Partly supportedcites [86] Hu, 2016, Epigenetic regulation of interleukin 6 by histone acetylation in macrophages and its role in paraquat-induced pulmonary fibrosis, Front. Immunol. sourceExperimental — accuracy not yet measured
The Discussion reports that histone modifications at the IL-6 promoter regulate its transcriptional activity, supporting the epigenetic-regulation part of the sentence. The paper does not establish the listed TLR ligands, IL-1, TNFα, ROS, and zinc as inducers of IL-6 transcription. No page numbers are available in our copy. (LLM, conf=0.86)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Could not verify
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Further, they act on fibroblasts and endothelial cells to recruit T helper 17 (Th17) cells, leading to an RA-like joint disease (F759 arthritis) and experimental autoimmune encephalomyelitis (EAE) (54, 66, 99).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Partly supportedcites [54] Arima, 2012, Regional neural activation defines a gateway for autoreactive T cells to cross the blood-brain barrier, Cell sourceExperimental — accuracy not yet measured
The Introduction (page unavailable) links IL-6 amplifier activation to F759 arthritis and EAE, and the Results (page unavailable) show that endothelial-cell IL-6 signalling contributes to pathogenic CD4+ T-cell accumulation and EAE severity. The paper also discusses Th17-cell recruitment in joints, but the passages do not establish a role for sIL-6Rα or show that fibroblasts specifically mediate the recruitment claimed here. (LLM, conf=0.86)
- Partly supportedcites [66] Ogura, 2008, Interleukin-17 promotes autoimmunity by triggering a positive-feedback loop via interleukin-6 induction, Immunity sourceExperimental — accuracy not yet measured
The Introduction and Discussion link IL-6 signaling in fibroblasts to F759 arthritis, and the EAE experiments report that disrupting IL-6–STAT3 signaling in type I collagen-positive cells attenuated Th17-cell-mediated EAE. Our copy does not establish that IL-6 together with soluble IL-6Rα acts on endothelial cells to recruit Th17 cells; it reports soluble IL-6R measurement, not that recruitment mechanism. (LLM, conf=0.85)
- Partly supportedcites [99] Sawa, 2006, Autoimmune arthritis associated with mutated interleukin (IL)-6 receptor gp130 is driven by STAT3/IL-7-dependent homeostatic proliferation of CD4+ T cells, J. Exp. Med. sourceExperimental — accuracy not yet measured
The Results (page not provided) show that IL-6 plus soluble IL-6 receptor induces IL-7 expression in fibroblasts, and that IL-6 contributes to RA-like arthritis in F759 mice. The paper does not establish that fibroblasts or endothelial cells recruit Th17 cells, or link this mechanism to EAE. (LLM, conf=0.88)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“The gp130-mediated activation of STAT3 is involved in the cell-cycle transition through the up-regulation of cyclins D2, D3 and A, and cdc25A, and the concomitant down-regulation of p21 and p27, while p21 is induced when STAT3 activation is suppressed, indicating the balance of contradictory signals determines the final output (110).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
What the sources say
- Supportedcites [110] Fukada, 1998, STAT3 orchestrates contradictory signals in cytokine-induced G1 to S cell-cycle transition, EMBO J. source
Split into 2 atoms — 2 confirmed, 0 partially supported, 0 disputed, 0 misattributed, 0 unverifiable. Expand the claim to see per-atom verdicts.
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“In addition to the aforementioned role of IL-6 on Tfh cells, IL-6 is known to direct the differentiation of IL-4-producing T helper type 2 (Th2) cells, while it inhibits the differentiation of IFNγ-producing Th1 cells (155).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
What the sources say
- Supportedcites [155] Rincón, 1997, Interleukin (IL)-6 directs the differentiation of IL-4-producing CD4+ T cells, J. Exp. Med. source
The Results and Discussion report that IL-6 directed CD4+ cells toward an IL-4-producing Th2 phenotype and impaired IL-12-driven Th1 differentiation, with reduced IFN-γ production. The paper presents these findings from its T-cell differentiation experiments; the supplied passages do not give page numbers. (LLM, conf=0.94)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“Much evidence has accumulated to show that STAT3 functions as an inhibitor for DC differentiation, maturation and antigen-presenting ability, and therefore STAT3 inhibits tumor immunity by interfering with DC functions (Fig. 2) (25).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
What the sources say
- Supportedcites [25] Rebe, 2019, STAT3, a master regulator of anti-tumor immune response, Cancer source
Section 3.2 (Dendritic Cells) states that STAT3 activation impairs DC antigen presentation and that IL-6–STAT3 signaling suppresses DC maturation and can dampen DC differentiation. It also reports that removing STAT3 enhances DC maturation and antitumor immunity in a melanoma model. The supplied passages do not specify page numbers. (LLM, conf=0.92)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The sources with a verdict support this claim; the rest weren’t checked yet, have no full text we could read, or their verdicts aren’t rated yet.
In the paper
“Taken together with the fact that the development of pristane or mineral oil-induced plasmacytoma and obesity-related hepatocellular carcinoma (HCC), both of which are related to inflammation, are dependent on IL-6 (254, 258), all studies indicate the intimate link between inflammation and tumorigenesis via IL-6–STAT3 signaling.”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Not checked — no full text of the sourcecites [254] Lattanzio, 1997, Defective development of pristane-oil-induced plasmacytomas in interleukin-6-deficient BALB/c mice, Am. J. Pathol.
- Supportedcites [258] Park, 2010, Dietary and genetic obesity promote liver inflammation and tumorigenesis by enhancing IL-6 and TNF expression, Cell source
The Results section, “Enhanced IL-6 production is required for obesity-induced tumor promotion,” reports that obese IL-6-deficient mice showed no significant increase in HCC tumor load. This supports the obesity-related HCC portion attributed to this paper; the separate plasmacytoma claim is paired with another citation. (LLM, conf=0.90)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“By utilizing IL-6-deficient mice, it was shown that IL-6 is required for type II collagen-induced arthritis (249), antigen-induced arthritis (AIA) (250), EAE (251, 252), pristane-induced autoantibody production (253) and plasmacytoma (254)”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Supportedcites [249] Alonzi, 1998, Interleukin 6 is required for the development of collagen-induced arthritis, J. Exp. Med. source
The citation to Alonzi et al. (249) is attached specifically to type II collagen-induced arthritis, not to the other conditions in the sentence. In the Discussion, the paper reports that genetically ablating IL-6 resulted in total protection from collagen-induced arthritis. (LLM, conf=0.94)
- Partly supportedcites [250] Ohshima, 1998, Interleukin 6 plays a key role in the development of antigen-induced arthritis, Proc. Natl Acad. Sci. USA sourceExperimental — accuracy not yet measured
Citation (250) is attached to antigen-induced arthritis (AIA), not the other conditions in the sentence. The Results and Discussion report that IL-6-deficient mice developed AIA, but it was markedly less severe than in wild-type mice. IL-6 is therefore important for severe AIA, but the paper does not show that it is required for AIA to occur. (LLM, conf=0.90)
- Not checked — no full text of the sourcecites [251] Eugster, 1998, IL-6-deficient mice resist myelin oligodendrocyte glycoprotein-induced autoimmune encephalomyelitis, Eur. J. Immunol. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“Further, IL-6–STAT3 signaling inhibits the LPS-induced maturation of DCs and DC-mediated activation of T cells in vivo (204).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
What the sources say
- Supportedcites [204] Park, 2004, IL-6 regulates in vivo dendritic cell differentiation through STAT3 activation, J. Immunol. source
The section on DC differentiation following LPS stimulation reports that LPS-induced maturation was enhanced in IL-6 knockout mice and suppressed in mice with enhanced gp130-mediated STAT3 signaling. The section on the in vivo T-cell response reports greater DC-dependent T-cell activation in IL-6 knockout mice and less in the gp130 mutant mice. Together, these findings support the cited paraphrase. (LLM, conf=0.91)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“Even in cancer development, it is now known that only 5–10% of all cancers are due to an inherited gene defect, and the remaining 90–95% have their roots in an environment and lifestyle that are intimately related with chronic inflammation (233).”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Supportedcites [233] Anand, 2008, Cancer is a preventable disease that requires major lifestyle changes, Pharm. Res. source
The Introduction states that only 5–10% of cancers are due to inherited gene defects, and Prevention of Cancer assigns the remaining 90–95% to environment and lifestyle. The Conclusions propose that cancer-causing lifestyle factors are linked through chronic inflammation and state that chronic inflammation is closely linked to tumorigenesis. (LLM, conf=0.92)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“The requirement of IL-6 and STAT3 for activated mutant EGFR-induced lung cancer and colitis-associated cancer has also been demonstrated (18, 257).”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Supportedcites [18] Grivennikov, 2009, IL-6 and Stat3 are required for survival of intestinal epithelial cells and development of colitis-associated cancer, Cancer Cell source
The Discussion reports that IL-6 and STAT3 signaling have a direct role in colitis-associated colon carcinogenesis and that removing STAT3 from intestinal epithelial cells interferes with tumor formation and growth in a mouse model. The sentence has two citations; this paper supports the colitis-associated cancer part, not the separately cited lung-cancer part. Page numbers were not available in our copy. (LLM, conf=0.92)
- Supportedcites [257] Gao, 2007, Mutations in the EGFR kinase domain mediate STAT3 activation via IL-6 production in human lung adenocarcinomas, J. Clin. Invest. source
For the lung-cancer part of the sentence, the Results report that reducing IL-6 inhibited STAT3 phosphorylation and growth of mutant-EGFR lung cancer cell lines both in vitro and in vivo. The paper’s concluding passage also states that IL-6 or JAK blockade inhibited growth and tumorigenesis. The separate colitis-associated-cancer part is not attributed to this source alone. (LLM, conf=0.86)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“IL-6 and IL-10 are abundant in tumor microenvironments originated from myeloid cells, fibroblasts and tumor cells, and they further induce M2 macrophage differentiation through STAT3 (215, 216).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
What the sources say
- Not checked — no full text of the sourcecites [215] Solís-Martínez, 2018, Regulation of immunophenotype modulation of monocytes-macrophages from M1 into M2 by prostate cancer cell-culture supernatant via transcription factor STAT3, Immunol. Lett. source
- Partly supportedcites [216] Fu, 2017, Interleukin 6 induces M2 macrophage differentiation by STAT3 activation that correlates with gastric cancer progression, Cancer Immunol. Immunother. sourceExperimental — accuracy not yet measured
The section ‘IL-6 induces M2 macrophage differentiation via STAT3 phosphorylation’ shows that IL-6 induces M2 differentiation through STAT3 activation. The Discussion also says IL-10 can promote M2 differentiation, but does not establish that IL-10 does so through STAT3. Our copy reports elevated IL-6 in gastric tumors; it does not establish that both cytokines are abundant across tumor microenvironments or identify myeloid cells, fibroblasts, and tumor cells as their sources. No page numbers are available in our copy. (LLM, conf=0.88)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Accordingly, IL-6 transgenic mice develop polyclonal plasmacytosis and monoclonal plasmacytoma in C57BL/6 and BALB/c genetic backgrounds, respectively (147, 246).”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Not checked — no full text of the sourcecites [147] Suematsu, 1989, IgG1 plasmacytosis in interleukin 6 transgenic mice, Proc. Natl Acad. Sci. USA source
- Partly supportedcites [246] Suematsu, 1992, Generation of plasmacytomas with the chromosomal translocation t(12;15) in interleukin 6 transgenic mice, Proc. Natl Acad. Sci. USA sourceExperimental — accuracy not yet measured
The Abstract reports massive plasmacytosis in C57BL/6-origin IL-6 transgenic mice and monoclonal plasmacytomas after introduction of a BALB/c genetic background. It does not establish that the C57BL/6 plasmacytosis was polyclonal. (LLM, conf=0.82)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Although the development of F759 arthritis is dependent on Th17 cells, the F759 mutation is required in non-immune cells but not immune cells, clearly indicating that the interaction between non-immune cells and immune cells is crucial for arthritis development in F759 mice (66, 99).”
§ IL-6 in inflammation, autoimmunity and cancer > Cross-talk between non-immune cells and immune cells through the IL-6 Amp
What the sources say
- Partly supportedcites [66] Ogura, 2008, Interleukin-17 promotes autoimmunity by triggering a positive-feedback loop via interleukin-6 induction, Immunity sourceExperimental — accuracy not yet measured
The Introduction states that the F759 mutation in nonhematopoietic cells is essential for arthritis development and that the mutation is not required in T cells. It does not establish that the mutation is unnecessary in all immune cells, as the claim says. No page number is available in our copy. (LLM, conf=0.86)
- Supportedcites [99] Sawa, 2006, Autoimmune arthritis associated with mutated interleukin (IL)-6 receptor gp130 is driven by STAT3/IL-7-dependent homeostatic proliferation of CD4+ T cells, J. Exp. Med. source
The Results section on IL-6, CD4+ T cells, and the gp130 mutation (page not provided) reports that arthritis developed when nonhematopoietic cells carried F759, regardless of whether transplanted bone marrow carried it. It did not develop in wild-type hosts even when the bone marrow carried F759. The paper concludes that the mutation was necessary in nonhematopoietic cells but not in CD4+ T cells. (LLM, conf=0.94)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“In addition, the enhanced production of epiregulin and TNFα is induced by the activation of the IL-6 Amp, and these molecules further activate NF-κB alone or STAT3 as well (10).”
§ IL-6 in inflammation, autoimmunity and cancer > Cross-talk between non-immune cells and immune cells through the IL-6 Amp
What the sources say
- Partly supportedcites [10] Murakami, 2013, Disease-association analysis of an inflammation-related feedback loop, Cell Rep. sourceExperimental — accuracy not yet measured
The Results report that epiregulin expression increases after IL-6 and IL-17 stimulation and that epiregulin enhances NF-κB activation through the PI3Kα pathway. The available passages do not support the parallel claims about TNFα production or TNFα activating NF-κB or STAT3; searches for TNF and tumor necrosis factor found no relevant passage. Page numbers were not provided. (LLM, conf=0.86)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Of note, the local low-grade inflammation that characterizes aging is triggered by senescent cells and is intimately related to age-related diseases such as cardiovascular diseases and cancer (5, 291, 292).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [5] Campisi, 2005, Senescent cells, tumor suppression, and organismal aging: good citizens, bad neighbors, Cell sourceExperimental — accuracy not yet measured
In the passage headed “Bad Neighbors—Senescent Cells Alter Tissue Structure and Function” (indexed as Abstract; page not provided), the paper says senescent fibroblasts secrete inflammatory cytokines and that local inflammation is frequent in aging tissues and proposed as a factor in atherosclerosis and cancer. It says senescent cells *might contribute* to local inflammation, not that they definitively trigger it; the citing sentence overstates that causal link. (LLM, conf=0.88)
- Not checked — no full text of the sourcecites [291] Lasry, 2015, Senescence-associated inflammatory responses: aging and cancer perspectives, Trends Immunol. source
- Partly supportedcites [292] Franceschi, 2017, Inflammaging and ‘Garb-aging’, Trends Endocrinol. Metab. sourceExperimental — accuracy not yet measured
The section “Aging is characterized by chronic low-grade inflammation” links inflammaging to age-related diseases, including cardiovascular disease and cancer. Box 4 says senescent cells may contribute to chronic inflammation through their pro-inflammatory secretions, while “Where do the stimuli fuelling inflammaging come from?” emphasizes cellular debris and altered self-molecules. The claim overstates the evidence by saying the inflammation is triggered by senescent cells. (LLM, conf=0.88)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“The disease induction requires IL-17A production by transferred T cells, IL-6 and CCL20 expression and STAT3 signaling in type I collagen-expressing cells.”
§ IL-6 in inflammation, autoimmunity and cancer > The local initiation model
What the sources say
- Partly supportedcites [52] Murakami, 2011, Local microbleeding facilitates IL-6- and IL-17-dependent arthritis in the absence of tissue antigen recognition by activated T cells, J. Exp. Med. sourceExperimental — accuracy not yet measured
The Results report that IL-6-deficient mice and mice lacking STAT3 in type I collagen-expressing cells did not develop arthritis, and that joint CCL20 knockdown prevented arthritis after Th17 transfer and microbleeding. But the Results also report mild arthritis after transfer of IL-17A-deficient Th17 cells. IL-17A therefore contributes to disease induction but is not strictly required as the claim states. (LLM, conf=0.90)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“This effect induces the manifestation of inflammatory diseases and autoimmune diseases (232).”
§ IL-6 in inflammation, autoimmunity and cancer > The local initiation model
What the sources say
- Partly supportedcites [232] Murakami, 2011, A four-step model for the IL-6 amplifier, a regulator of chronic inflammations in tissue-specific MHC class II-associated autoimmune diseases, Front. Immunol. sourceExperimental — accuracy not yet measured
The Abstract (page unavailable) says chronic IL-6 amplifier activation leads to the manifestation of autoimmune diseases in the paper’s tissue-specific model. The section “The IL-6 Amplifier Beyond MHC Class II-Associated Diseases and Disorders” discusses a possible role in certain other chronic inflammatory diseases, but does not establish the claim for inflammatory diseases generally. (LLM, conf=0.83)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Shi et al. showed that FFA activates TLR4 signaling and that mice lacking TLR4 are partially protected against high fat diet-induced insulin resistance (306).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [306] Shi, 2006, TLR4 links innate immunity and fatty acid-induced insulin resistance, J. Clin. Invest. sourceExperimental — accuracy not yet measured
The section on high-fat diets and the Figure 8 caption report partial protection against insulin resistance in female TLR4-knockout mice; the Discussion reports no corresponding insulin-sensitivity difference in males. The paper also reports that FFAs activate TLR4 signaling. The claim omits the female-specific qualification and thus implies a broader finding than the paper supports. (LLM, conf=0.88)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“The enlargement of adipose tissue in obesity induces mechanical stress, ER stress and hypoxia in adipocytes, resulting in the release of free fatty acids (FFAs) and inflammatory cytokines such as IL-6 and TNFα and chemokines (303–305), suggesting activation of the IL-6 Amp.”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [303] Ghosh, 2013, An IL-6 link between obesity and cancer, Front. Biosci. (Elite Ed.) sourceExperimental — accuracy not yet measured
Split into 2 atoms — 0 confirmed, 2 partially supported, 0 disputed, 0 misattributed, 0 unverifiable. Expand the claim to see per-atom verdicts.
- Partly supportedcites [304] Kern, 2019, Obesity-induced TNFalpha and IL-6 signaling: the missing link between obesity and inflammation-driven liver and colorectal cancers, Cancers sourceExperimental — accuracy not yet measured
Split into 2 atoms — 0 confirmed, 2 partially supported, 0 disputed, 0 misattributed, 0 unverifiable. Expand the claim to see per-atom verdicts.
- Partly supportedcites [305] Trayhurn, 2008, Hypoxia in adipose tissue: a basis for the dysregulation of tissue function in obesity?, Br. J. Nutr. sourceExperimental — accuracy not yet measured
Split into 2 atoms — 0 confirmed, 2 partially supported, 0 disputed, 0 misattributed, 0 unverifiable. Expand the claim to see per-atom verdicts.
hedgeddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Obesity causes chronic low-grade inflammation and is closely related with chronic inflammatory diseases, such as cardiovascular diseases and T2D and some cancers, such as breast, liver and colon cancers (233, 302–304).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [233] Anand, 2008, Cancer is a preventable disease that requires major lifestyle changes, Pharm. Res. sourceExperimental — accuracy not yet measured
The Obesity section (page unavailable) links obesity with inflammation and reports associations with colon, liver, and breast cancer mortality, specifying postmenopausal women for breast cancer. The Conclusions (page unavailable) also identify obesity as a risk factor that activates an inflammatory pathway. Our copy does not establish that obesity causes chronic low-grade inflammation or link it specifically to cardiovascular disease or type 2 diabetes. (LLM, conf=0.86)
- Not checked — no full text of the sourcecites [302] Haslam, 2005, Obesity, Lancet source
- Supportedcites [303] Ghosh, 2013, An IL-6 link between obesity and cancer, Front. Biosci. (Elite Ed.) source
The Introduction links obesity to cardiovascular disease, type 2 diabetes and cancer; section 4.1 describes obesity-associated chronic low-grade inflammation. The paper’s table of obesity-related cancers lists colorectal, breast and liver cancers (with breast cancer specified as postmenopausal). (LLM, conf=0.91)
- Partly supportedcites [304] Kern, 2019, Obesity-induced TNFalpha and IL-6 signaling: the missing link between obesity and inflammation-driven liver and colorectal cancers, Cancers sourceExperimental — accuracy not yet measured
Section 1, “Obesity-Induced Inflammation Impairs Insulin Action” (page unavailable), describes obesity-driven chronic low-grade inflammation and its effects on insulin sensitivity. Section 7, “Obesity-Induced Inflammation and CRC” (page unavailable), reports a strong association between obesity and colorectal cancer; the paper also discusses liver cancer. The passages do not establish the claimed associations with cardiovascular disease or breast cancer, and they address insulin resistance rather than directly establishing the T2D association. (LLM, conf=0.85)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“IL-6–STAT3 signaling in adipocytes is known to be involved in adipocyte-induced EMT in breast cancer cells (23) and obesity-promoted HCC (258).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Supportedcites [23] Gyamfi, 2018, Interleukin-6/STAT3 signalling regulates adipocyte induced epithelial-mesenchymal transition in breast cancer cells, Sci. Rep. source
The Introduction reports that human adipocytes induce an EMT phenotype in breast cancer cells through paracrine IL-6/STAT3 signalling. The Results locate STAT3 activation in the co-cultured breast cancer cells, rather than in adipocytes. The HCC clause has a separate citation (258) and is not attributed to this source. (LLM, conf=0.92)
- Partly supportedcites [258] Park, 2010, Dietary and genetic obesity promote liver inflammation and tumorigenesis by enhancing IL-6 and TNF expression, Cell sourceExperimental — accuracy not yet measured
The Results section, “Enhanced IL-6 production is required for obesity-induced tumor promotion,” reports that IL-6 deficiency prevented STAT3 activation in obese mouse livers and HCCs and largely eliminated obesity-associated tumor promotion. The Discussion places IL-6–STAT3 signaling in hepatocytes, not adipocytes; it does not establish that signaling within adipocytes promotes HCC. The breast-cancer portion is attached to a different citation and is not assessed here. (LLM, conf=0.88)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Consistently, these events recruit macrophages, which further produce IL-6 and TNFα, major players involved in obesity-induced low-grade inflammation and cancer development (303, 304).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [303] Ghosh, 2013, An IL-6 link between obesity and cancer, Front. Biosci. (Elite Ed.) sourceExperimental — accuracy not yet measured
Section 4.1 (page unavailable) says adipose-tissue hypoxia can recruit macrophages and that obese adipose tissue secretes IL-6 and TNF-α, contributing to inflammation. Section 4.2 links these cytokines to cancer-related processes. The cited passages do not specifically establish that the recruited macrophages themselves produce both cytokines in this obesity-related sequence. (LLM, conf=0.82)
- Supportedcites [304] Kern, 2019, Obesity-induced TNFalpha and IL-6 signaling: the missing link between obesity and inflammation-driven liver and colorectal cancers, Cancers source
Section 1 describes immune-cell recruitment to obese white adipose tissue and identifies infiltrating M1 macrophages as a main source of TNFα and IL-6 in obesity-induced low-grade inflammation. The same section states that these cytokines also promote cancer. (LLM, conf=0.93)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Concerning age-related diseases, senescent cells themselves are critically involved in T1D and T2D, and the deletion of senescent beta cells prevents diabetes (57, 298), indicating that even in T1D in which tissue-specific T cells are involved (299–301), local initiators originating from senescent beta cells play critical roles.”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Not checked — no full text of the sourcecites [57] Thompson, 2019, Targeted elimination of senescent beta cells prevents type 1 diabetes, Cell Metab. source
- Partly supportedcites [298] Aguayo-Mazzucato, 2019, Acceleration of β cell aging determines diabetes and senolysis improves disease outcomes, Cell Metab. sourceExperimental — accuracy not yet measured
The Discussion (page unavailable) reports that deleting p16Ink4a-positive cells improved glucose homeostasis and β-cell function in mouse models of aging and insulin resistance, supporting a role for senescent β-cells in T2D-related dysfunction. It does not show that deletion prevents diabetes. For T1D, the Discussion describes senolysis as a possible application and cites Thompson et al. (2019) for evidence of senescent β-cells in a T1D mouse model; it does not establish a critical role in T1D. (LLM, conf=0.89)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
Every source the paper cites for this claim was checked and supports it.
In the paper
“Indeed, Chang et al. showed that IL-6–STAT3 signaling is increased at the leading edge of breast tumors to promote MDSC recruitment, angiogenesis, fibroblast infiltration and metastases (83).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Supportedcites [83] Chang, 2013, The IL-6/JAK/Stat3 feed-forward loop drives tumorigenesis and metastasis, Neoplasia source
The Discussion (page not supplied) reports that IL-6 and pStat3 were highest at the invasive edge of mammary tumors, alongside abundant fibroblasts, endothelial cells, and MDSCs. It also reports that IL-6 increased these stromal cells, while another Discussion passage links IL-6/Stat3 signaling to angiogenesis and metastatic potential. (LLM, conf=0.93)
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Both hepatitis B and hepatitis C viruses, which induce hepatitis and cause hepatocellular carcinoma, activate STAT3 (328–330), while HTLV-I, Epstein–Barr virus and papilloma virus activate STAT3 and NF-κB (311, 331, 332).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Not checked — no full text of the sourcecites [311] Migone, 1995, Constitutively activated Jak-STAT pathway in T cells transformed with HTLV-I, Science source
- Partly supportedcites [331] Muromoto, 2009, Epstein-Barr virus-derived EBNA2 regulates STAT3 activation, Biochem. Biophys. Res. Commun. sourceExperimental — accuracy not yet measured
The Introduction reports STAT3 activation in HTLV-I-transformed T cells and Epstein–Barr virus-related lymphoma cell lines. A title in the References mentions NF-κB activation induced by an Epstein–Barr virus protein, but the paper does not address papilloma virus or establish the claim that all three viruses activate both STAT3 and NF-κB. No page numbers are available in our copy. (LLM, conf=0.84)
- Not checked — no full text of the sourcecites [332] Sun, 2002, PTEN is a negative regulator of STAT3 activation in human papillomavirus-infected cells, J. Gen. Virol. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Indeed, a human colonic bacterium, enterotoxigenic Bacteroides fragilis (ETBF)-derived toxin, which triggers colitis and induces colonic tumors in mice, activates the STAT3 and Th17 response (326).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [326] Wu, 2009, A human colonic commensal promotes colon tumorigenesis via activation of T helper type 17 T cell responses, Nat. Med. sourceExperimental — accuracy not yet measured
The Results report that ETBF colonization caused colitis and colonic tumors in Min mice and activated colonic Stat3 and TH17 responses. The Discussion suggests that ETBF’s toxin, BFT, has a central role, but the reported mouse findings concern colonization by the toxin-producing bacterium, not an isolated-toxin experiment; the claim attributes those effects more directly to the toxin than the paper establishes. (LLM, conf=0.86)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“The loss of p53 alone is insufficient to initiate intestinal tumorigenesis but markedly enhances carcinogen-induced tumor incidence, the progression of which is associated with increased intestinal permeability, causing the formation of a NF-κB-dependent inflammatory microenvironment (322).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Supportedcites [322] Schwitalla, 2013, Loss of p53 in enterocytes generates an inflammatory microenvironment enabling invasion and lymph node metastasis of carcinogen-induced colorectal tumors, Cancer Cell source
Split into 2 atoms — 2 confirmed, 0 partially supported, 0 disputed, 0 misattributed, 0 unverifiable. Expand the claim to see per-atom verdicts.
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
Every source the paper cites for this claim was checked and supports it.
In the paper
“Mutated K-RAS activates NF-κB in a mouse model of lung cancer (315), and the K-RAS-induced activation of NF-κB is required for pancreatic cancer development (316), which depends on IL-6 trans-signaling-mediated STAT3 activation (317).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Supportedcites [316] Ling, 2012, KrasG12D-induced IKK2/β/NF-κB activation by IL-1α and p62 feedforward loops is required for development of pancreatic ductal adenocarcinoma, Cancer Cell source
The Discussion states that constitutive NF-κB activation is required for pancreatic ductal adenocarcinoma development and describes its activation by KrasG12D. The IL-6/STAT3 clause has a separate citation (317) and is not attributed to this source. (LLM, conf=0.94)
- Supportedcites [317] Lesina, 2011, Stat3/Socs3 activation by IL-6 transsignaling promotes progression of pancreatic intraepithelial neoplasia and development of pancreatic cancer, Cancer Cell source
The citation attaches to the IL-6 trans-signaling–mediated STAT3 activation clause, not the NF-κB requirement attributed to citation 316. The Discussion states that KrasG12D-induced pancreatic oncogenesis involves IL-6 trans-signaling and STAT3, and that disrupting them impedes PanIN progression and PDAC development. (LLM, conf=0.93)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The sources with a verdict support this claim; the rest weren’t checked yet, have no full text we could read, or their verdicts aren’t rated yet.
In the paper
“Additionally, either or both are activated in cells transformed with HTLV-1 (311), v-src (312), abl (313), bcr-abl or v-eyk (314).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Not checked — no full text of the sourcecites [311] Migone, 1995, Constitutively activated Jak-STAT pathway in T cells transformed with HTLV-I, Science source
- Not checked — no full text of the sourcecites [312] Yu, 1995, Enhanced DNA-binding activity of a Stat3-related protein in cells transformed by the Src oncoprotein, Science source
- Supportedcites [313] Hilbert, 1996, Distinct tumorigenic potential of abl and raf in B cell neoplasia: abl activates the IL-6 signaling pathway, Immunity source
Citation 313 is attached to abl, not the other listed transforming agents. The paper reports in its Results that Stat3 was constitutively phosphorylated in abl/myc tumor cell lines, supporting activation of STAT3 in those cells. (LLM, conf=0.92)
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Either or both of STAT3 and NF-κB are activated in a variety of tumors, including lung, breast, pancreatic and prostate cancers (67).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [67] Grivennikov, 2010, Dangerous liaisons: STAT3 and NF-kappaB collaboration and crosstalk in cancer, Cytokine Growth Factor Rev. sourceExperimental — accuracy not yet measured
The STAT3 section (page unavailable) reports activated STAT3 in breast, lung and prostate cancers, and the Introduction describes NF-κB and STAT3 activation across malignancies. Our copy does not identify pancreatic cancer among the examples, so that part of the attributed list is unsupported (searched: pancreatic). (LLM, conf=0.85)
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Helicobacter pylori, which induces gastritis and is related to gastric cancer, activates STAT3 in epithelial cells (327).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [327] Bronte-Tinkew, 2009, Helicobacter pylori cytotoxin-associated gene A activates the signal transducer and activator of transcription 3 pathway in vitro and in vivo, Cancer Res. sourceExperimental — accuracy not yet measured
The Abstract reports STAT3 activation in H. pylori-infected epithelial cells, and the Discussion identifies H. pylori as a major risk factor for gastric cancer. Our copy discusses gastric inflammation but does not explicitly state that H. pylori induces gastritis, so that part of the sentence is not verified. (LLM, conf=0.83)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Both hepatitis B and hepatitis C viruses, which induce hepatitis and cause hepatocellular carcinoma, activate STAT3 (328–330), while HTLV-I, Epstein–Barr virus and papilloma virus activate STAT3 and NF-κB (311, 331, 332).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Partly supportedcites [328] Yoshida, 2002, Activation of STAT3 by the hepatitis C virus core protein leads to cellular transformation, J. Exp. Med. sourceExperimental — accuracy not yet measured
The Results section (‘The HCV-Core Protein Induces Activation of STAT3’; page not available) reports STAT3 activation by HCV core protein and stronger activation with the full-length HCV genome. The Introduction mentions HBV proteins only in a comparison of other signaling assays; our copy does not show that HBV activates STAT3. Thus, this source supports the HCV part, not the HBV part. (LLM, conf=0.88)
- Not checked — no full text of the sourcecites [329] Choudhari, 2007, Deactivation of Akt and STAT3 signaling promotes apoptosis, inhibits proliferation, and enhances the sensitivity of hepatocellular carcinoma cells to an anticancer agent, Atiprimod, Mol. Cancer Ther. source
- Partly supportedcites [330] Xiong, 2017, Hepatitis B virus promotes cancer cell migration by downregulating miR-340-5p expression to induce STAT3 overexpression, Cell Biosci. sourceExperimental — accuracy not yet measured
The Background (page unavailable) associates both HBV and HCV infection with hepatocellular carcinoma. The Results (page unavailable) report that HBV increases STAT3 expression and that phosphorylated STAT3 follows the same trend. The paper does not report HCV activating STAT3 or establish that either virus induces hepatitis. (LLM, conf=0.88)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“ARDS in COVID-19 is considered the result of cytokine storm (12–17), resembling the disease caused by SARS-CoV-1 and Middle East respiratory syndrome coronavirus (MERS-CoV) (335).”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Partly supportedcites [12] Mehta, 2020, COVID-19: consider cytokine storm syndromes and immunosuppression, Lancet sourceExperimental — accuracy not yet measured
The main text (page not supplied) says a subgroup of patients with severe COVID-19 might have a cytokine storm syndrome and identifies ARDS as a leading cause of mortality. It does not establish that COVID-19 ARDS generally results from a cytokine storm. The SARS/MERS comparison has a separate citation in the citing sentence and is not judged here. (LLM, conf=0.84)
- Supportedcites [13] Mahmudpour, 2020, COVID-19 cytokine storm: the anger of inflammation, Cytokine source
The citation is attached to the comparison with SARS-CoV and MERS-CoV. The Introduction reports immune involvement in severe infections caused by all three coronaviruses and increased Th17 responses in each, supporting the claimed resemblance in inflammatory disease. (LLM, conf=0.86)
- Not checked — no full text of the sourcecites [14] Hu, 2021, The cytokine storm and COVID-19, J. Med. Virol. source
- Supportedcites [15] Hirano, 2020, COVID-19: a new virus, but a familiar receptor and cytokine release syndrome, Immunity source
The citation to this source is attached to the comparison with SARS-CoV-1 and MERS-CoV, not to the separate cytokine-storm clause. The Main Text states that severe ARDS is a major phenotype of COVID-19, just as in disease caused by SARS-CoV and MERS-CoV. No page number is provided in our copy. (LLM, conf=0.94)
- Supportedcites [16] García, 2020, Immune response, inflammation, and the clinical spectrum of COVID-19, Front. Immunol. source
The citation is attached to the comparison with SARS and MERS, not to the preceding cytokine-storm claim. The Histopathology section states that post-mortem findings in COVID-19 are similar to those reported in SARS and MERS. (LLM, conf=0.90)
- Supportedcites [17] Vabret, 2020, Immunology of COVID-19: current state of the science, Immunity source
Citation (335) is attached to the comparison with SARS-CoV-1 and MERS-CoV, not to the preceding claim that cytokine storm causes ARDS. The Concluding Remarks state that severe COVID-19 pathology resembles certain immunopathologies seen in those infections, including cytokine release syndrome. (LLM, conf=0.92)
- Supportedcites [335] de Wit, 2016, SARS and MERS: recent insights into emerging coronaviruses, Nat. Rev. Microbiol. source
Citation (335) is attached to the comparison with SARS-CoV and MERS-CoV, not to the preceding claim that a cytokine storm causes ARDS in COVID-19. In “The pathogenesis of SARS-CoV and MERS-CoV,” the paper reports ARDS in both diseases and associates progression to ARDS with elevated pro-inflammatory cytokines and chemokines. (LLM, conf=0.87)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“The AngII–AT1R signaling system plays an important role not only in pneumonia but also in ARDS model mice (345), indicating that lung injury itself activates the AngII–AT1R signaling.”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Partly supportedcites [345] Imai, 2005, Angiotensin-converting enzyme 2 protects from severe acute lung failure, Nature sourceExperimental — accuracy not yet measured
The Main section reports that AngII acting through the AT1a receptor has a causative role in acute lung failure in mice, supporting the ARDS-model portion of the claim. The Methods describe acid-aspiration, sepsis and endotoxin models; although the Main section names pneumonia as a condition whose lung pathology these models may mimic, it does not separately demonstrate an AngII–AT1R role in pneumonia. Our copy provides no page numbers for these passages. (LLM, conf=0.85)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Since an activated IL-6 Amp enhances the production of chemokines to recruit lymphoid cells and myeloid cells including Th17 cells in the lesion, the IL-6 Amp could be further enhanced (66).”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Partly supportedcites [66] Ogura, 2008, Interleukin-17 promotes autoimmunity by triggering a positive-feedback loop via interleukin-6 induction, Immunity sourceExperimental — accuracy not yet measured
The paper reports that IL-17A and IL-6 synergistically increase expression of KC and MIP2 in fibroblasts (results passage labeled Abstract; no page available), and it proposes an IL-6 signaling amplifier involving Th17 cells (Abstract; no page available). It does not establish that these chemokines recruit lymphoid and myeloid cells, including Th17 cells, into lesions or that such recruitment further enhances the amplifier. (LLM, conf=0.85)
hedgeddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“This axis activates ADAM17, which generates the mature form of EGFR ligand and TNFα, two NF-κB stimulators (346).”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Partly supportedcites [346] Eguchi, 2018, Understanding angiotensin II type 1 receptor signaling in vascular pathophysiology, Hypertension sourceExperimental — accuracy not yet measured
The ADAM17 section states that AT1R activates ADAM17, which generates a mature EGFR ligand, and identifies TNFα as another ADAM17 substrate. It does not identify those two products as NF-κB stimulators; the ER Stress section instead links NF-κB activation to sustained ER stress. Page numbers are unavailable in our copy. (LLM, conf=0.88)
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
The cited sources that could be checked couldn’t confirm or dispute this claim.
In the paper
“Furthermore, ADAM17 increases sIL-6Rα, which activates STAT3 through gp130 on a variety of non-immune cells including fibroblasts, endothelial cells and epithelial cells (33, 96).”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Could not verifycites [33] Murakami, 2019, Pleiotropy and specificity: insights from the interleukin 6 family of cytokines, Immunity sourceExperimental — accuracy not yet measured
The supplied passages and additional searches for ADAM17, sIL-6R, gp130, STAT3, and fibroblasts returned only reference links, with no section or page text addressing the claim. Our copy therefore provides no passage by which to assess the proposed signaling mechanism or the listed cell types. (LLM, conf=0.76)
- Not checked — no full text of the sourcecites [96] Rose-John, 2017, The soluble interleukin 6 receptor: advanced therapeutic options in inflammation, Clin. Pharmacol. Ther. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Could not verify
Every source the paper cites for this claim was checked and supports it.
In the paper
“Therefore, it is likely that the AngII–AT1R axis is involved in the ARDS induced by SARS-CoV-2 infection (15), which is like SARS-CoV-1-induced ARDS in a mouse model (344).”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Supportedcites [15] Hirano, 2020, COVID-19: a new virus, but a familiar receptor and cytokine release syndrome, Immunity source
The Main Text proposes that dysregulation of the AngII–AT1R pathway after SARS-CoV-2 infection could contribute to inflammatory disease and ARDS. This supports the citing sentence’s hedged claim that the axis is likely involved. The mouse-model comparison is attached to a different citation and is not judged here. (LLM, conf=0.92)
- Supportedcites [344] Kuba, 2005, A crucial role of angiotensin converting enzyme 2 (ACE2) in SARS coronavirus-induced lung injury, Nat. Med. source
The citation to this paper is attached to the comparison with SARS-CoV-induced lung injury in a mouse model, not to the preceding SARS-CoV-2 proposition. In the Main section, SARS-CoV Spike increased lung AngII in injured mice, and AT1R inhibition attenuated their severe lung injury and pulmonary edema. Page numbers are not provided in our copy. (LLM, conf=0.90)
hedgedconditional- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Supported
The cited sources support only part of this claim — read each source’s verdict below.
In the paper
“Baricitinib, an inhibitor of JAK1 and JAK2, is reported to prevent COVID-19 severity followed by lowering IL-6 and TNFα levels (369).”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Partly supportedcites [369] Bronte, 2020, Baricitinib restrains the immune dysregulation in patients with severe COVID-19, J. Clin. Invest. sourceExperimental — accuracy not yet measured
The Introduction identifies baricitinib as a JAK1/JAK2 inhibitor, and the section “Baricitinib affects the immune landscape in COVID-19 patients” reports reduced IL-6 and TNFα after treatment. But preventing progression to severe COVID-19 was a hypothesis, not an established result: the clinical-parameters section reports improved oxygenation and an association with lower mortality, but no significant difference in ARDS incidence. Page numbers are not available in our copy. (LLM, conf=0.88)
hedgeddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
- Partly supported
A claim is checked only against the cited paper’s full text, never its abstract, and none is available for its sources.
In the paper
“Studies in the 1980s also showed that IL-6 is produced by cardiac myxoma cells (42) and Castleman’s germinal center cells (43)”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [42] Hirano, 1987, Human B-cell differentiation factor defined by an anti-peptide antibody and its possible role in autoantibody production, Proc. Natl Acad. Sci. USA source
- Not checked — no full text of the sourcecites [43] Yoshizaki, 1989, Pathogenic significance of interleukin-6 (IL-6/BSF-2) in Castleman’s disease, Blood source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it. It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“The original hypothesis has since been updated and is here called the local initiation model, which emphasizes the importance of local initiators, such as infection, injury and stress in the promotion of interactions between non-immune cells and immune cells in disease.”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
In the paper
“It was originally identified as B-cell stimulating factor-2 (BSF-2), an inducer of immunoglobulin production (27).”
§ IL-6 in inflammation, autoimmunity and cancer > Introduction
What the sources say
- Not checked — no full text of the sourcecites [27] Hirano, 1986, Complementary DNA for a novel human interleukin (BSF-2) that induces B lymphocytes to produce immunoglobulin, Nature source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“The trans-signaling mode expands the range of cells that respond to IL-6, because gp130 is expressed in a wide range of cells, including endothelial cells and fibroblasts, explaining one of the mechanisms by which IL-6 acts on a wide range of cells and tissues to exert a pleiotropic function.”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
In the paper
“It was shown that myeloid-derived sIL-6R complexed with IL-6 inhibits Th1 responses through STAT3-induced c-Maf. This is considered to be one of mechanisms by which IL-6 inhibits T-cell immunity against tumors (156).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
What the sources say
- Not checked — no full text of the sourcecites [156] Tsukamoto, 2017, Soluble IL6R expressed by myeloid cells reduces tumor-specific Th1 differentiation and drives tumor progression, Cancer Res. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
None of the sources cited for this claim have been checked yet.
In the paper
“IL-6 activates Janus kinase (JAK) family tyrosine kinases through gp130, leading to the activation of STAT family transcription factors, mainly STAT3, as well as src homology region 2 domain-containing phosphatase 2 (SHP-2), GRB2-associated-binding protein (GAB), extracellular signal-regulated kinase (ERK) and mitogen-activated protein kinase (MAPK) signaling pathways (Fig. 1A).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A claim is checked only against the cited paper’s full text, never its abstract, and none is available for its sources.
In the paper
“IL-6 transgenic mice show severe plasmacytosis with hypergammaglobulinemia (147), while IL-6-deficient mice are defective in immunoglobulin production (148, 149).”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
What the sources say
- Not checked — no full text of the sourcecites [147] Suematsu, 1989, IgG1 plasmacytosis in interleukin 6 transgenic mice, Proc. Natl Acad. Sci. USA source
- Not checked — no full text of the sourcecites [148] Kopf, 1994, Impaired immune and acute-phase responses in interleukin-6-deficient mice, Nature source
- Not checked — no full text of the sourcecites [149] Ramsay, 1994, The role of interleukin-6 in mucosal IgA antibody responses in vivo, Science source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A claim is checked only against the cited paper’s full text, never its abstract, and none is available for its sources.
In the paper
“Another knock-in mouse strain (gp130Y757F/Y757F mice) having the point mutation Y759F in murine gp130 also shows the hyper-activation of STAT3 and develops gastric hyperplasia resembling human early-stage gastric cancer (64, 256).”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Not checked — no full text of the sourcecites [64] Jenkins, 2005, Hyperactivation of Stat3 in gp130 mutant mice promotes gastric hyperproliferation and desensitizes TGF-beta signaling, Nat. Med. source
- Not checked — no full text of the sourcecites [256] Tebbutt, 2002, Reciprocal regulation of gastrointestinal homeostasis by SHP2 and STAT-mediated trefoil gene activation in gp130 mutant mice, Nat. Med. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
In the paper
“In fact, it takes about 1 year for F759 mice to develop arthritis, while gp130Y757F/Y757F mice show hyperproliferative lesions in the stomach mucosa as early as 6 weeks (256).”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Not checked — no full text of the sourcecites [256] Tebbutt, 2002, Reciprocal regulation of gastrointestinal homeostasis by SHP2 and STAT-mediated trefoil gene activation in gp130 mutant mice, Nat. Med. source
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“Moreover, IL-6 itself is an autocrine and paracrine growth factor as well as a survival factor for a variety of cancers and promotes cancer development and progression by inducing angiogenesis and metastasis.”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: unclear — It is unclear why the paper says it holds.
It may state two separate claims as one.
In the paper
“Furthermore, F759 arthritis is enhanced by human T-cell lymphotropic virus 1 (HTLV-1) pX gene, which encodes Tax, a transcriptional trans-activator of NF-κB, showing the synergistic effect of STAT3 and NF-κB (255).”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Not checked — no full text of the sourcecites [255] Ishihara, 2004, The point mutation of tyrosine 759 of the IL-6 family cytokine receptor gp130 synergizes with HTLV-1 pX in promoting rheumatoid arthritis-like arthritis, Int. Immunol. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Of note is that F759 mice spontaneously develop IL-6-dependent and CD4(+) T-cell-dependent F759 arthritis in a manner dependent on age, supporting the idea that enhanced but dysregulated IL-6–STAT3 signaling is involved in age-related inflammatory autoimmune diseases, such as RA.”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
hedgeddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
In the paper
“In addition, serum IL-6 levels are increased and correlate with the poor prognosis of multiple myeloma patients (235).”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
What the sources say
- Not checked — no full text of the sourcecites [235] Ludwig, 1991, Interleukin-6 is a prognostic factor in multiple myeloma, Blood source
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Thus, it seems the IL-6–STAT3 axis is involved in inflammation, autoimmunity and cancer and that they are all intimately related to each other.”
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
hedgedcorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
In the paper
“It is also consistent with the fact that many chronic inflammatory diseases are dependent on aging and accompanied by the activation of polyclonal T cells and an increase of IL-6 (285).”
§ IL-6 in inflammation, autoimmunity and cancer > The local initiation model
What the sources say
- Not checked — no full text of the sourcecites [285] Ershler, 1993, Interleukin-6 and aging: blood levels and mononuclear cell production increase with advancing age and in vitro production is modifiable by dietary restriction, Lymphokine Cytokine Res.
plaincorrelational- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Thus, the IL-6 Amp plays a crucial role in not only chronic inflammatory and autoimmune diseases but also cancer.”
§ IL-6 in inflammation, autoimmunity and cancer > Cross-talk between non-immune cells and immune cells through the IL-6 Amp
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“The IL-6 Amp can become dysregulated depending on the genetic background and other factors including local initiators, leading to inflammatory diseases and autoimmune diseases.”
§ IL-6 in inflammation, autoimmunity and cancer > The local initiation model
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A claim is checked only against the cited paper’s full text, never its abstract, and none is available for its sources.
In the paper
“In line with these findings, FFA activates NF-κB through TLR4, leading to the expression of monocyte chemotactic protein-1 (MCP-1) to recruit macrophages, which further produce IL-6 and TNFα (307, 308).”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
What the sources say
- Not checked — no full text of the sourcecites [307] Davis, 2008, Tlr-4 deficiency selectively protects against obesity induced by diets high in saturated fat, Obesity (Silver Spring) source
- Not checked — no full text of the sourcecites [308] Schaeffler, 2009, Fatty acid-induced induction of Toll-like receptor-4/nuclear factor-kappaB pathway in adipocytes links nutritional signalling with innate immunity, Immunology source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“The IL-6 Amp enhances the production of a variety of pro-inflammatory cytokines and chemokines, resulting in the recruitment of immune cells, such as activated T cells into the lesion.”
§ IL-6 in inflammation, autoimmunity and cancer > The local initiation model
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: unclear — It is unclear why the paper says it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“These studies provide evidence supporting the claim that a variety of stressors, such as gravity and pain, act as local initiators through specific neural pathways, resulting in activation of the IL-6 Amp, linking inflammation and autoimmunity and possibly affecting cancer development and progression.”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
hedgeddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: unclear — It is unclear why the paper says it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“TLR stimulation also induces IL-6 production and other pro-inflammatory cytokines through NF-κB, leading to activation of the IL-6 Amp.”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“Infection is a local initiator that triggers and maintains inflammation, leading to chronic inflammatory diseases and cancer.”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“All the above results are in line with cancer cells activating the IL-6 Amp through the direct or indirect activation of NF-κB and STAT3, likely resulting in the recruitment of Th17 cells and/or myeloid cells to further enhance the IL-6 Amp.”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
hedgedconditional- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“Intracellular S1P induces IL-6 production through NF-κB activation, resulting in the activation of both STAT3 and NF-κB, major players of the IL-6 Amp.”
§ IL-6 in inflammation, autoimmunity and cancer > Local initiators activating the IL-6 Amp in chronic inflammation
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: unclear — It is unclear why the paper says it holds.
It may state two separate claims as one.
In the paper
“Furthermore, it also occurs due to lung damage caused by gastric acid or inappropriate operation of a ventilator, like aspiration pneumonia (334).”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
What the sources say
- Not checked — no full text of the sourcecites [334] Thompson, 2017, Acute respiratory distress syndrome, N. Engl. J. Med. source
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: unclear — It is unclear whether the paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“IL-6 usually acts on lymphoid and myeloid immune cells expressing both IL-6Rα and gp130, but it acts on a wide range of non-immune cells as well, including fibroblasts and endothelial cells that express gp130 during inflammation.”
§ IL-6 in inflammation, autoimmunity and cancer > Conclusion and perspectives
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“As discussed in this manuscript, these risk factors are all associated with chronic inflammation, which could decrease the threshold of the activation of the IL-6 Amp to induce cytokine storm.”
§ IL-6 in inflammation, autoimmunity and cancer > Cytokine storm, uncontrolled acute inflammation
hedgedconditional- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: fails — The paper’s conclusions may not depend on it.
- Based: fails — The paper may give no reason why it holds.
Probably not a substantial claim. The paper’s conclusions may not depend on it. The paper may give no reason why it holds. It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“One possible explanation for why inhibiting IL-6 activity is effective against inflammation-related diseases, in spite of the many cytokines involved, is that in inflammation IL-6 is the major stimulator of STAT3, which plays important roles in inflammation and oncogenesis together with NF-κB, which expresses IL-6 as a target.”
§ IL-6 in inflammation, autoimmunity and cancer > Conclusion and perspectives
hedgeddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: unclear — It is unclear why the paper says it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“STAT3 and NF-κB are involved in a wide range of inflammatory diseases and cancer development and, more importantly, their interaction is central to activation of the IL-6 Amp.”
§ IL-6 in inflammation, autoimmunity and cancer > Conclusion and perspectives
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: unclear — It is unclear why the paper says it holds.
It may state two separate claims as one.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“It has since been clarified that IL-6 is a critical factor in inflammation, autoimmunity and cancer, and its inhibitors are used as therapeutic agents for several diseases already, with the potential of being expanded to more in the future.”
§ IL-6 in inflammation, autoimmunity and cancer > Conclusion and perspectives
boosteddirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The paper states this without a source or data behind it here, so there is nothing to check it against. The quote shows it is in the paper.
In the paper
“Several inhibitors of IL-6 are now clinically used, but in order to develop small peptides or compounds with higher specificity and efficacy against diseases, high-resolution structural analysis of the ligand–receptor complex is needed.”
§ IL-6 in inflammation, autoimmunity and cancer > Conclusion and perspectives
plain- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
Probably not a substantial claim.
A conclusion the paper draws from its own results — it cites no other source for it, so there is none to check. The quote shows it is in the paper.
In the paper
“A variety of local initiators, such as infection, stressors, injury, senescence, obesity, pre-neoplastic mutation, cell death and smoking can activate the IL-6 Amp, which mediates the interaction between non-immune cells and immune cells, leading to the manifestation of inflammatory and autoimmune diseases and cancer.”
§ IL-6 in inflammation, autoimmunity and cancer > Conclusion and perspectives
plaindirect- Asserted: passes — The authors put it forward as true.
- Checkable: passes — It can be checked against evidence.
- Load-bearing: passes — The paper’s conclusions depend on it.
- Based: passes — The paper gives a reason why it holds.
It may state two separate claims as one.
The authors state this as an assumption.
The same cause can induce both RA and IL-6 expression.
“Assuming that the same cause can induce both RA and IL-6 expression, such an intimate relationship could be explained.”
Self-check
Percentages, sums and reported differences re-computed from the paper’s own numbers.
Each contradiction between two passages listed here was confirmed by an independent check.
Rated minor: it would not touch a result — a value no result rests on, a p-value that stays on the same side of the significance level, or a conflict that is clerical, resolved elsewhere in the paper, or outside its own findings.
In the paper
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
“IL-6/STAT3 induces the differentiation of M2 macrophages and activates MDSCs, both of which have anti-tumor immunity effects.”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6–STAT3 signaling in immunity
“IL-6–STAT3 signaling induces the conversion of Treg cells to Th17 cells, which are pro-inflammatory, and induces the differentiation of M2 macrophages and MDSCs, both of which suppress tumor immunity.”
Rated minor: it would not touch a result — a value no result rests on, a p-value that stays on the same side of the significance level, or a conflict that is clerical, resolved elsewhere in the paper, or outside its own findings.
In the paper
§ IL-6 in inflammation, autoimmunity and cancer > Involvement of IL-6 in chronic inflammation, autoimmune diseases and cancer
“Another knock-in mouse strain (gp130Y757F/Y757F mice) having the point mutation Y759F in murine gp130”
§ IL-6 in inflammation, autoimmunity and cancer > IL-6 expression, IL-6 receptor and its mode of action and signaling
“Phosphorylated Y759 (human gp130) or Y757 (murine gp130) serves as a binding site for SOCS3 and SHP-2.”
Citations
Cites
Clinical features of patients infected with 2019 novel coronavirus in Wuhan, China
Huang, C., Wang, Y., Li, X., Ren, L., Zhao, J., Hu, Y., Zhang, L., Fan, G., Xu, J., Gu, X., et al. (2020) · The Lancet · doi:10.1016/s0140-6736(20)30183-5 · Cited by 48,884 · cited 11×
“ARDS in COVID-19 is considered the result of cytokine storm (12–17), re-sembling the disease caused by SARS-CoV-1 and Middle East respiratory syndrome coronavirus (MERS-CoV) (335).”
BackgroundInfluentialZhou, F., Yu, T., Du, R., Fan, G., Liu, Y., Liu, Z., Xiang, J., Wang, Y., Song, B., Gu, X., et al. (2020) · The Lancet · doi:10.1016/s0140-6736(20)30566-3 · Cited by 27,263 · cited 14×
“ARDS in COVID-19 is considered the result of cytokine storm (12–17), re-sembling the disease caused by SARS-CoV-1 and Middle East respiratory syndrome coronavirus (MERS-CoV) (335).”
BackgroundInfluentialA pneumonia outbreak associated with a new coronavirus of probable bat origin
Zhou, P., Yang, X.-L., Wang, X.-G., Hu, B., Zhang, L., Zhang, W., Si, H.-R., Zhu, Y., Li, B., Huang, C.-L., et al. (2020) · Nature · doi:10.1038/s41586-020-2012-7 · Cited by 21,969
Chen, N., Zhou, M., Dong, X., Qu, J., Gong, F., Han, Y., Qiu, Y., Wang, J., Liu, Y., Wei, Y., et al. (2020) · The Lancet · doi:10.1016/s0140-6736(20)30211-7 · Cited by 21,415
Hoffmann, M., Kleine-Weber, H., Schroeder, S., Krüger, N., Herrler, T., Erichsen, S., Schiergens, T. S., Herrler, G., Wu, N.-H., Nitsche, A., et al. (2020) · Cell · doi:10.1016/j.cell.2020.02.052 · Cited by 20,286 · cited 1×
“Since SARS-CoV-2 uses angiotensin-converting enzyme (ACE) 2 as a receptor (342, 343), it is speculated that SARS-CoV-2 induces pro-inflammatory cytokines through angiotensin II (AngII)/angiotensin type 1 receptor (AT1R) (15) as SARS-CoV-1 does (344–346).”
BackgroundCoussens, L. M., & Werb, Z. (2002) · Nature · doi:10.1038/nature01322 · Cited by 15,111 · cited 2×
“Chronic inflammation is often accompanied with auto-immune disease and cancer (2, 4, 9).”
BackgroundDexamethasone in Hospitalized Patients with Covid-19
The RECOVERY Collaborative Group (2021) · New England Journal of Medicine · doi:10.1056/nejmoa2021436 · Cited by 11,477
Mantovani, A., Allavena, P., Sica, A., & Balkwill, F. (2008) · Nature · doi:10.1038/nature07205 · Cited by 11,399
Immunity, Inflammation, and Cancer
Grivennikov, S. I., Greten, F. R., & Karin, M. (2010) · Cell · doi:10.1016/j.cell.2010.01.025 · Cited by 10,755 · cited 3×
“Chronic inflammation is often accompanied with auto-immune disease and cancer (2, 4, 9).”
BackgroundHigh-performance medicine: the convergence of human and artificial intelligence
Topol, E. J. (2019) · Nature Medicine · doi:10.1038/s41591-018-0300-7 · Cited by 10,450 · cited 1×
“Improvements in detection technology and systems biology research for age-and inflammation-related diseases driven by artificial intelligence (AI) would be helpful (375).”
Background
Cited by
Sorted by citations
Interleukins in cancer: from biology to therapy
Briukhovetska, D., Dörr, J., Endres, S., Libby, P., Dinarello, C. A., & Kobold, S. (2021) · Nature reviews. Cancer · doi:10.1038/s41568-021-00363-z · Cited by 822
Free radicals and their impact on health and antioxidant defenses: a review
Chandimali, N., Bak, S. G., Park, E. H., Lim, H. J., Won, Y., Kim, E., Park, S., & Lee, S. (2025) · Cell Death Discovery · doi:10.1038/s41420-024-02278-8 · Cited by 852
M1/M2 macrophages and their overlaps – myth or reality?
Strizova, Z. O., Benešová, I., Bartolini, R., Novysedlák, R., Cecrdlova, E., Foley, L. K., & Stříž, I. (2023) · Clinical Science · doi:10.1042/cs20220531 · Cited by 709
Emerging role of exosomes in cancer progression and tumor microenvironment remodeling
Paskeh, M. D. A., Entezari, M., Mirzaei, S., Zabolian, A., Saleki, H., Naghdi, M. J., Sabet, S., Khoshbakht, M. A., Hashemi, M., Hushmandi, K., et al. (2022) · Journal of Hematology & Oncology · doi:10.1186/s13045-022-01305-4 · Cited by 591
Targeting cytokine and chemokine signaling pathways for cancer therapy
Yi, M., Li, T., Niu, M., Zhang, H., Wu, Y., Wu, K., & Dai, Z. (2024) · Signal Transduction and Targeted Therapy · doi:10.1038/s41392-024-01868-3 · Cited by 528
Metadata
- Authors
- Hirano, T.
- Affiliations
- Hokkaido University (JP)
- Corresponding author
- Hirano, Toshio
- Venue
- International Immunology
- Publisher
- Oxford University Press (OUP)
- Published
- 2020-12-18
- Volume
- 33
- Issue
- 3
- Pages
- 127-148
- ISSN
- 0953-8178 (ISSN-L)
- 1460-2377
- PMC
- PMC7799025
- OpenAlex
- W3112344668
- Semantic Scholar
- 53b0b4afcb2ca15e75cb05c20a38e26d39ab02f8
- Cited by
- 1,453
- References
- 390
- Citations by year
- 2026: 246 · 2025: 394 · 2024: 326 · 2023: 273 · 2022: 165 · 2021: 49
- FWCI
- 39.33
- Type
- journal article
- Language
- en
- Open access
- hybrid
- doi.org
- Licence
- Locations
- International Immunology (publishedVersion, cc-by-nc)
- PubMed (publishedVersion)
- Europe PMC (PubMed Central) (submittedVersion, cc-by-nc)
- PubMed Central (submittedVersion, cc-by-nc)
- QST-Repository (National Institutes for Quantum and Radiological Science and Technology) (submittedVersion, cc-by-nc-nd)
- Topics
- Life Sciences › Immunology and Microbiology › Immunology › Cytokine Signaling Pathways and Interactions
- Life Sciences › Biochemistry, Genetics and Molecular Biology › Cancer Research › NF-κB Signaling Pathways
- Health Sciences › Medicine › Rheumatology › Rheumatoid Arthritis Research and Therapies
- Keywords
- interleukin-6, chronic inflammation, autoimmunity, cancer, NF-κB, STAT3, cytokine storm, COVID-19 pandemic, senescence, obesity, positive feedback loop, hematopoiesis, bone metabolism
- MeSH terms
- COVID-19, COVID-19 Drug Treatment, Animals, Humans, Inflammation, Neoplasms, Autoimmunity, Interleukin-6, STAT3 Transcription Factor
About these checks
Registry, retraction, Cites and statistics results are computed, not judged, with one exception: a printed reference that no index lists is read once by a model before it counts as a citation error. Claims are extracted by a model and kept only when their quote appears verbatim in the text; each cited source is then checked, with a model’s help, for whether it says what the claim relies on. Those verdicts can be wrong — read the claim’s quote and the cited source before relying on one.
A language model drew the claims from the paper on 2026-10-23, under version 3 of the claim definition (model deepseek-v4.1-flash, prompt extract-v3).
A second model labelled each claim on 2026-10-23 (model jev-1.13-20260917, definition revision gold-guideline-v0.2).
Cited sources are checked with a language model’s help (gpt-6-sol).
Statistics and arithmetic are recomputed, not judged.
Confidence bands are shown only for fields calibrated on our annotated test set. None is calibrated yet, so no band is shown.
What counts as a claim ›How we measure confidence ›How we check this ›
Are you an author, or do you see a problem with this page? Request a review
